



Targeted mass spectrometry assays fail most often from transition design, cycle-time overload, or matrix interference—not from a mysterious instrument collapse. Multiple reaction monitoring (MRM) and parallel reaction monitoring (PRM) schedule closed peptides on liquid chromatography-tandem mass spectrometry (LC-MS/MS) (Lange et al. 2008; Peterson et al. 2012). In project language, weak or missing peaks, unstable ion ratios, missed retention windows, or global sensitivity loss after panel expansion are troubleshooting signals. Re-acquiring the same method without changing one locked factor usually reproduces the same failure. The guide below maps targeted-proteomics troubleshooting workflow, locked parameters, limits, and recovery fit. It is not a biomarker validation brochure, not a claim that discovery detectability guarantees targeted success, and not a license to change transitions, gradient, and platform in one unrecorded step.
Unmatched sample prep can mimic acquisition failure (Bantscheff et al. 2012). Relative fold change and amount with stable isotope-labeled internal standard (SIS) peptides remain separate claim units during recovery (Gerber et al. 2003).
In MtoZ Biolabs targeted offerings, failed assays are re-scoped after symptom pattern, matrix, panel size, platform, and quality control (QC) evidence are written in the brief. Service intake appears in Targeted Proteomics Service.
What Failure Signs Point to Which Layer
Four symptom layers belong in the troubleshooting record and stay separated.
Transition layer: weak or absent peaks in matrix despite neat standards; only one product ion reliable; unstable qualifier ratios.
Cycle-time layer: many targets lose sensitivity together after panel growth; scheduled windows look populated but peaks are undersampled.
Retention layer: targets fall outside scheduled windows between batches; standards and samples disagree on retention.
Matrix-and-prep layer: spike recovery collapses in plasma or tissue; replicate digests disagree before any schedule change.
Name the failing layer before editing the method. Mixed undiagnosed changes hide the root cause.
How Transitions, Cycle Time, and Matrix Interference Interact
The three title failure modes often stack.
Unstable or interference-prone transitions fail first in complex matrices even when literature product ions looked strong. Cycle-time overload appears when dwell × transition count no longer samples chromatographic peak width. Matrix interference can suppress ionization or contaminate low-resolution MRM channels; PRM fragment spectra may separate those cases when ion ratios stay unstable after transition redesign. Route choice after interference review appears in Parallel Reaction Monitoring vs MRM: Which Fits?.
Panel build and sample-count scoping before recovery appear in Scoping an MRM Quantitation Project: Transition Design, Sample Count, and Validation Deliverables. This page stays on failure diagnosis and recovery order.
End-to-End Troubleshooting Workflow
A practical recovery spine usually runs as follows.
- Capture chromatograms, transition or fragment lists, schedules, and QC from the failing batch.
- Classify the dominant symptom: missing peak, ratio instability, missed window, or global sensitivity loss.
- Re-test priority transitions or fragments in matrix-matched spikes versus neat standards—one change set at a time.
- Recalculate cycle time against peak width; cut or stage lower-priority analytes if undersampling is proven.
- Rebuild retention windows only after confirming whether drift or wrong scheduling caused misses.
- Check prep parity: salts, lipids, detergents, digest completeness, and load matching across arms.
- Move interference-limited peptides to PRM or alternate products when MRM ratios stay unstable after steps 3–6.
- Lock a revised method; run a small matrix pilot before the full cohort.
- Expand tier-two peptides only after tier-one pilots pass the declared rules.
Steps 1 through 2 diagnose. Steps 3 through 9 recover without rewriting the entire assay at once.
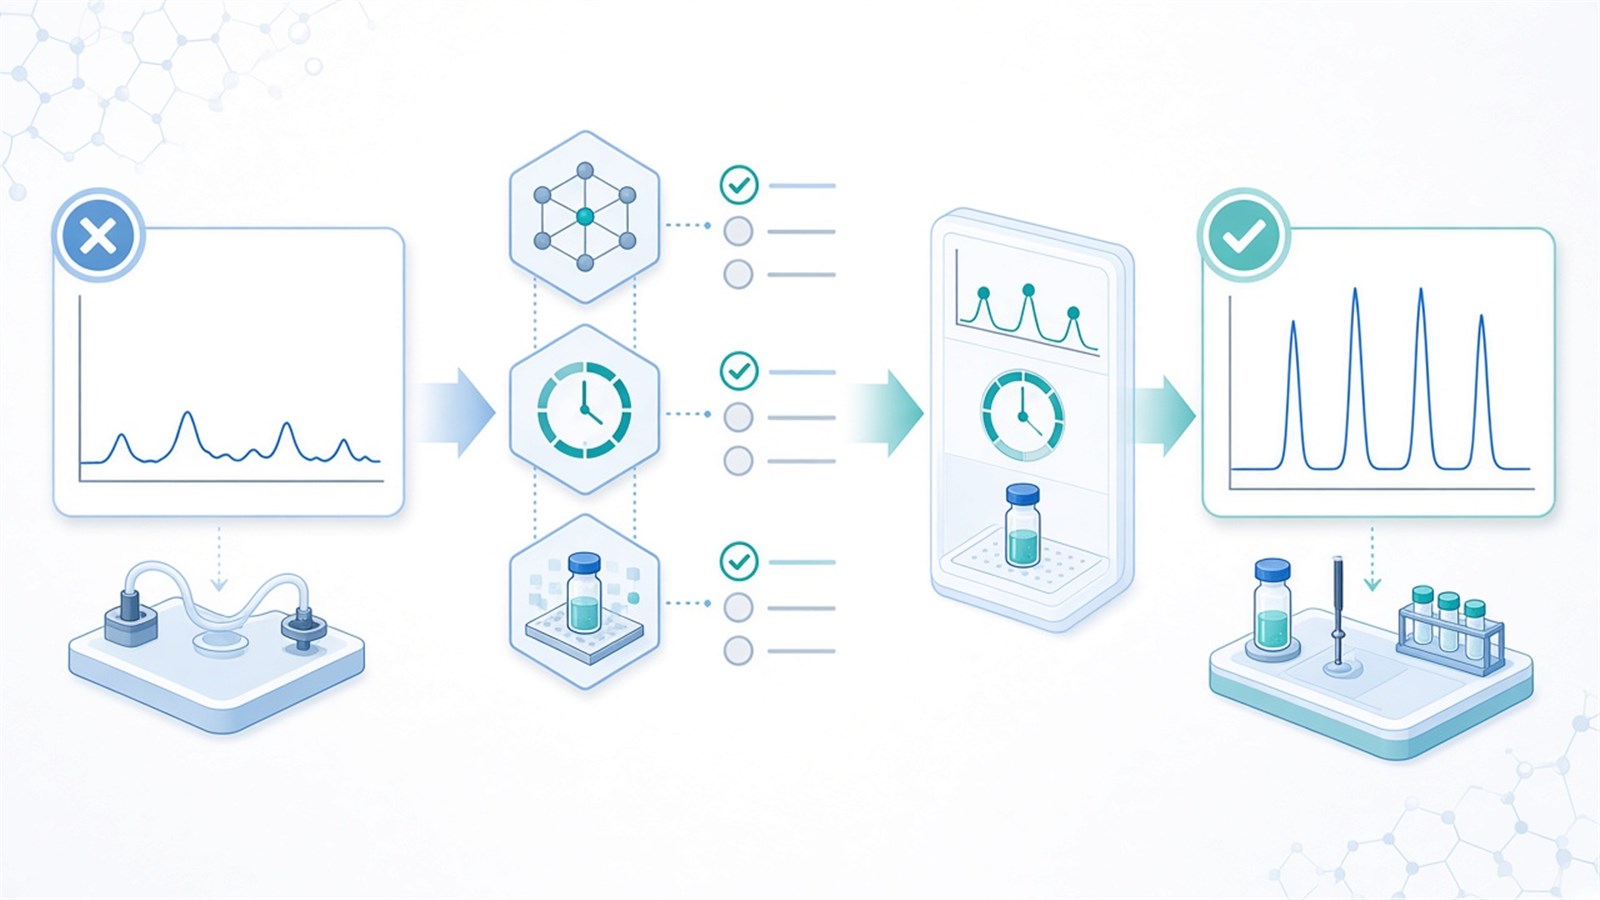
Figure 1. Fix one locked layer at a time: transitions, cycle time, retention, then matrix or platform.
Parameters to Re-Lock During Recovery
Layer |
Parameter to re-check |
Failure mode if skipped |
|---|---|---|
Symptom log |
Peak absence, ratio drift, missed window, global loss |
Wrong fix applied |
Transitions |
Precursor, products, collision energy |
Literature ions fail in matrix |
Qualifiers |
Ion-ratio rules |
Interference read as biology |
Cycle time |
Dwell × count vs peak width |
Panel-wide sensitivity loss |
Schedule |
Retention windows and drift tolerance |
Targets outside acquisition |
Prep |
Matched digest and cleanup |
Prep variance as assay failure |
Matrix spikes |
Neat vs matrix-matched response |
Suppression missed |
Platform |
MRM vs PRM for failing subset |
Forcing unstable MRM transitions |
Pilot n |
Small matrix pilot before cohort |
Full re-run of a broken method |
Deliverable |
Revised method and interference notes |
Same failure on resubmission |
Limits follow the same table. Troubleshooting does not create clinical or release decisions. Discovery detectability does not guarantee targeted matrix performance. Single-transition methods remain fragile when specificity is required.
How to Choose the Recovery Path
Use symptom class, panel pressure, and interference evidence together.
Observed problem |
Recovery path that often fits |
Recheck before committing |
|---|---|---|
Weak in matrix, strong in neat standard |
Transition redesign + matrix spikes |
Blind LC gradient rewrite |
Global sensitivity loss after panel growth |
Cut/stage panel; fix cycle time |
Add more transitions first |
Missed scheduled targets |
Retention rebuild after drift check |
Assume analyte absent |
Unstable MRM ion ratios in tissue/plasma |
Alternate products or PRM subset |
Keep one failing transition |
Digests disagree, QC instrument stable |
Prep parity repair |
Platform switch only |
Tier-one OK, tier-two weak |
Stage lower-priority method |
One oversized panel forever |
Method never piloted in study matrix |
Matrix pilot before cohort |
Immediate full re-acquisition |
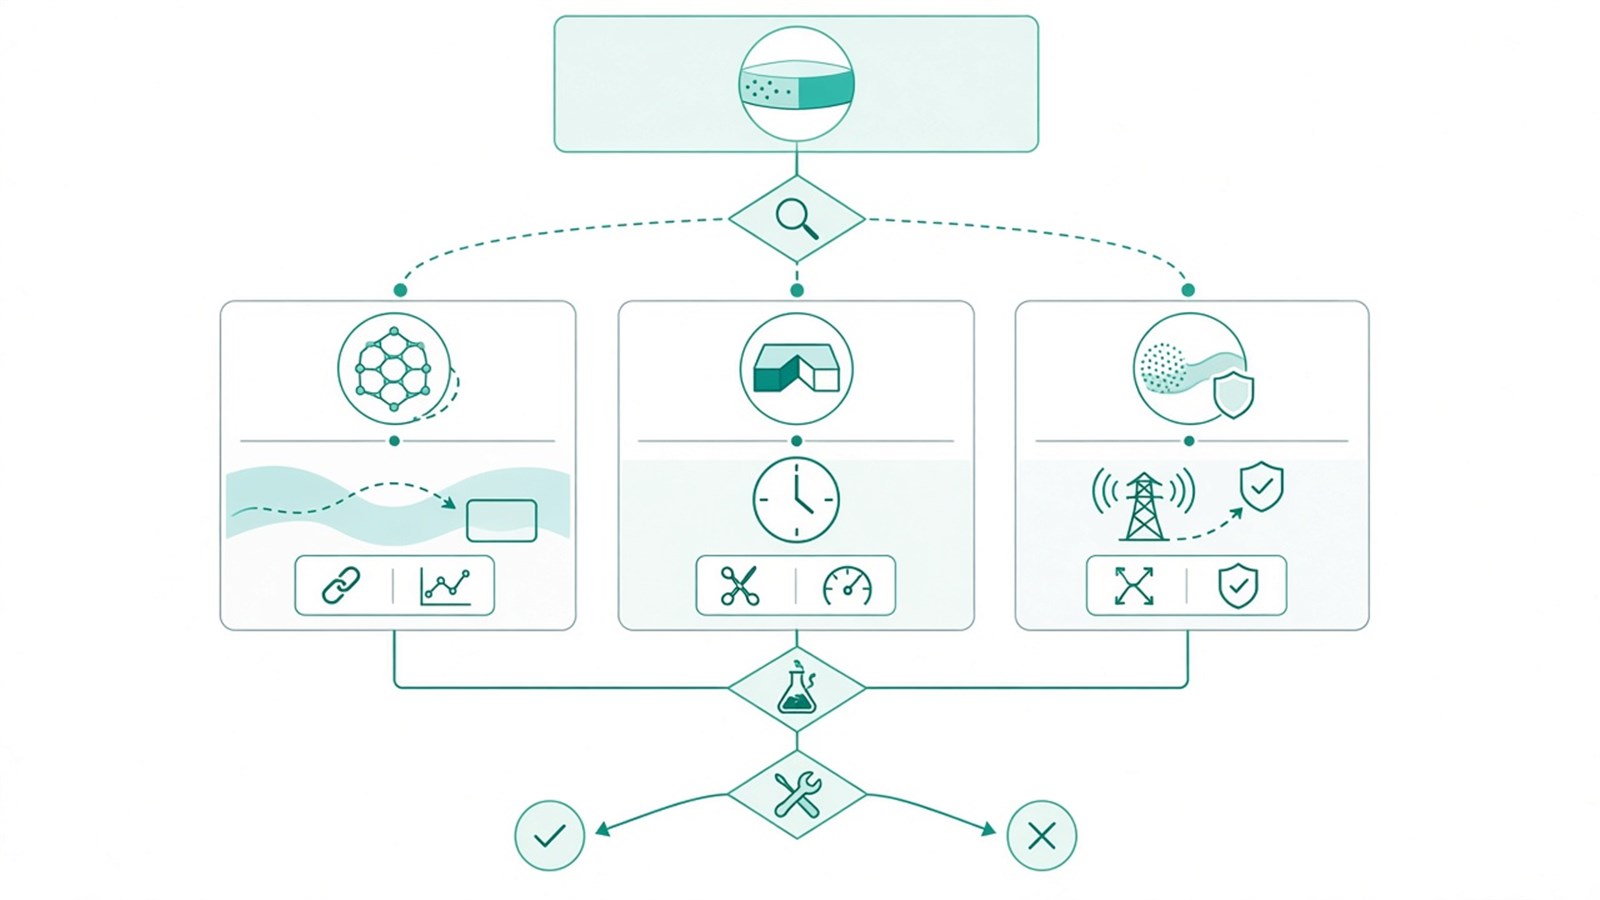
Figure 2. Match recovery to the failing layer; move to PRM or panel staging when interference or cycle time dominates.
MRM service intake appears in SRM-MRM Quantitative Proteomics Services. Teams with failing chromatograms, panel lists, and matrix notes can review MtoZ Biolabs targeted and PRM entries when those fields are declared in the recovery record.
Frequently Asked Questions
1. Should the same targeted method be re-run without changes?
Only after transitions, cycle time, retention, and matrix effects are reviewed. Repeating an unchanged failing method rarely helps.
2. Can PRM fix every failed MRM assay?
No. PRM helps when interference or fragment review is the limit. It does not repair unmatched prep or extreme cycle-time overload by itself.
3. What if only part of the panel works?
Keep tier-one peptides on the locked method and stage lower-priority analytes in a follow-on schedule.
4. Can sample prep alone fix weak targeted MS data?
Sometimes, when yield or cleanup is the dominant layer. Matrix pilots that compare neat versus matrix-matched spikes separate prep from transition failure.
5. What should be shared before a recovery review?
Analyte list, matrix, acquisition method, chromatograms, transition or fragment tables, QC behavior, and whether the claim is relative or SIS absolute.
MtoZ Biolabs can discuss targeted mass spectrometry troubleshooting when symptom class, panel status, and matrix evidence are documented.
If further evaluation of transition redesign, cycle-time cuts, or MRM-to-PRM fallback is needed, project details can be submitted for a feasibility review.
Reference
- V. Lange, P. Picotti, B. Domon, R. Aebersold (2008). Selected reaction monitoring for quantitative proteomics: a tutorial. Mol. Syst. Biol., 4, 222. https://doi.org/10.1038/msb.2008.61
- A.C. Peterson, J.D. Russell, A.A. Klammer, P.F. Doubleday, H. Liu, J.N. Anderson, M.J. MacCoss (2012). Parallel reaction monitoring for high resolution and high mass accuracy quantitative, targeted proteomics. Mol. Cell. Proteomics, 11, 1475-1488. https://doi.org/10.1074/mcp.O112.020271
- S.A. Gerber, J. Rush, O. Stemman, M.W. Kirschner, S.P. Gygi (2003). Absolute quantification of proteins and phosphoproteins from cell lysates by tandem MS. Proc. Natl. Acad. Sci. U.S.A., 100, 6940-6945. https://doi.org/10.1073/pnas.0832254100
- M. Bantscheff, S. Lemeer, H.U. Haab, B. Kuster (2012). Quantitative mass spectrometry in proteomics: a critical review. Anal. Bioanal. Chem., 404, 939-965. https://doi.org/10.1007/s00216-012-5892-4
