



Parallel reaction monitoring (PRM) is operated as a scheduled, targeted assay: finalize a proteotypic peptide list, digest and clean up the samples, optionally spike stable isotope-labeled standards (SIS), place precursor m/z and retention-time windows on an inclusion list, isolate each precursor, record a high-resolution fragment-ion spectrum, and quantify from extracted fragment peak areas in software such as Skyline. In scheduled PRM, predefined target precursors are monitored within specified retention-time windows. PRM does not discover an open proteome the way data-dependent acquisition (DDA) or data-independent acquisition (DIA) does. Relative comparison of the same peptide across samples is the usual result. Absolute amount requires a matched SIS added at a defined step.
If the peptide panel is already finalized, or the project is a targeted validation, the MtoZ Biolabs Parallel Reaction Monitoring (PRM) Service can review feasibility and assay planning for that project.
PRM Operational Workflow Step by Step
A workable PRM project follows a fixed order. Skipping an earlier step cannot be repaired by higher Orbitrap resolution alone.
1. Finalize the Target Peptide List
Targets usually come from a prior DDA or DIA discovery matrix, a pathway panel, or curated peptide resources. Prefer proteotypic peptides with stable chromatography and usable fragment ions. Shared peptides, heavily modified sequences, and poorly ionizing candidates should be flagged before the inclusion list is finalized.
2. Decide Relative Versus Absolute Readout
Same-peptide peak areas support relative comparison across groups when digestion and loading stay comparable. Absolute quantification requires an appropriate internal-standard and calibration strategy, with the SIS amount and spike timing documented for each quantified peptide. An SIS added after a failed digestion cannot reconstruct the lost protein.
3. Prepare Matched Digests
Protein extraction, reduction and alkylation, trypsin digestion, and peptide cleanup must keep sample loads comparable. Internal process standards can monitor handling drift. They do not convert a relative PRM assay into an absolute one unless the labeled analyte peptides are present.
4. Build the Acquisition Method
Schedule precursor m/z values and retention-time windows on a quadrupole-Orbitrap instrument. Isolation width, maximum fill time, resolution, and collision energy set how many precursors fit inside one chromatographic peak. Overpacking the schedule creates dropped points and unstable peak areas.
5. Acquire and Extract
Each scheduled precursor is isolated, fragmented, and recorded as a full high-resolution MS2 spectrum (Peterson et al. 2012). Quantitative and qualifier fragments are extracted as chromatograms. Peak area, retention-time alignment with the light or heavy pair, and fragment consistency are reviewed before a value enters the result table.
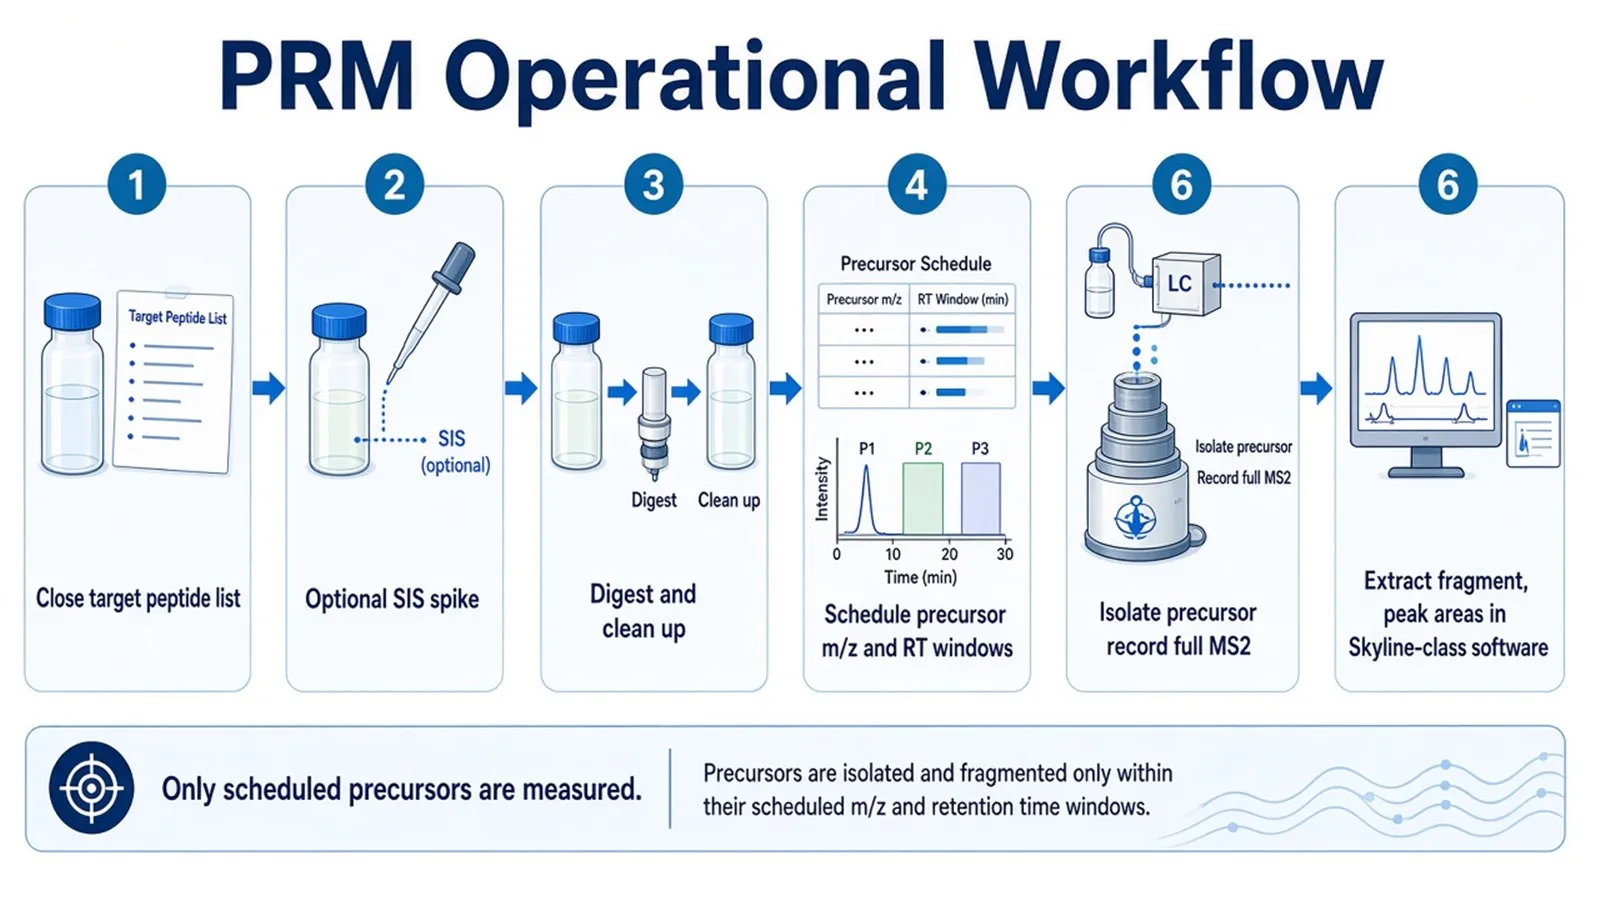
Figure 1. PRM is executed as a scheduled workflow: only listed precursors are fragmented, and quantification comes from extracted fragment peak areas.
If the precursor was never placed on the inclusion list, PRM will not measure it, regardless of abundance in the digest.
Principle: How One Scheduled Precursor Becomes a Peak Area
PRM uses a quadrupole to isolate a precursor window, fragments that ion population, and records product ions in parallel on a high-resolution analyzer (Peterson et al. 2012). Unlike multiple reaction monitoring (MRM) on a triple quadrupole, PRM does not require every quantitative transition to be chosen before acquisition. Fragments can be selected or replaced during review because the spectrum was stored.
The isolation window is still a window. Co-isolated ions can fragment with the target. Accurate-mass detection separates many interfering products; it cannot invent precursor uniqueness when overlapping fragments remain. That is why method records should state isolation width, resolution, and which fragments were used for quantification versus confirmation.
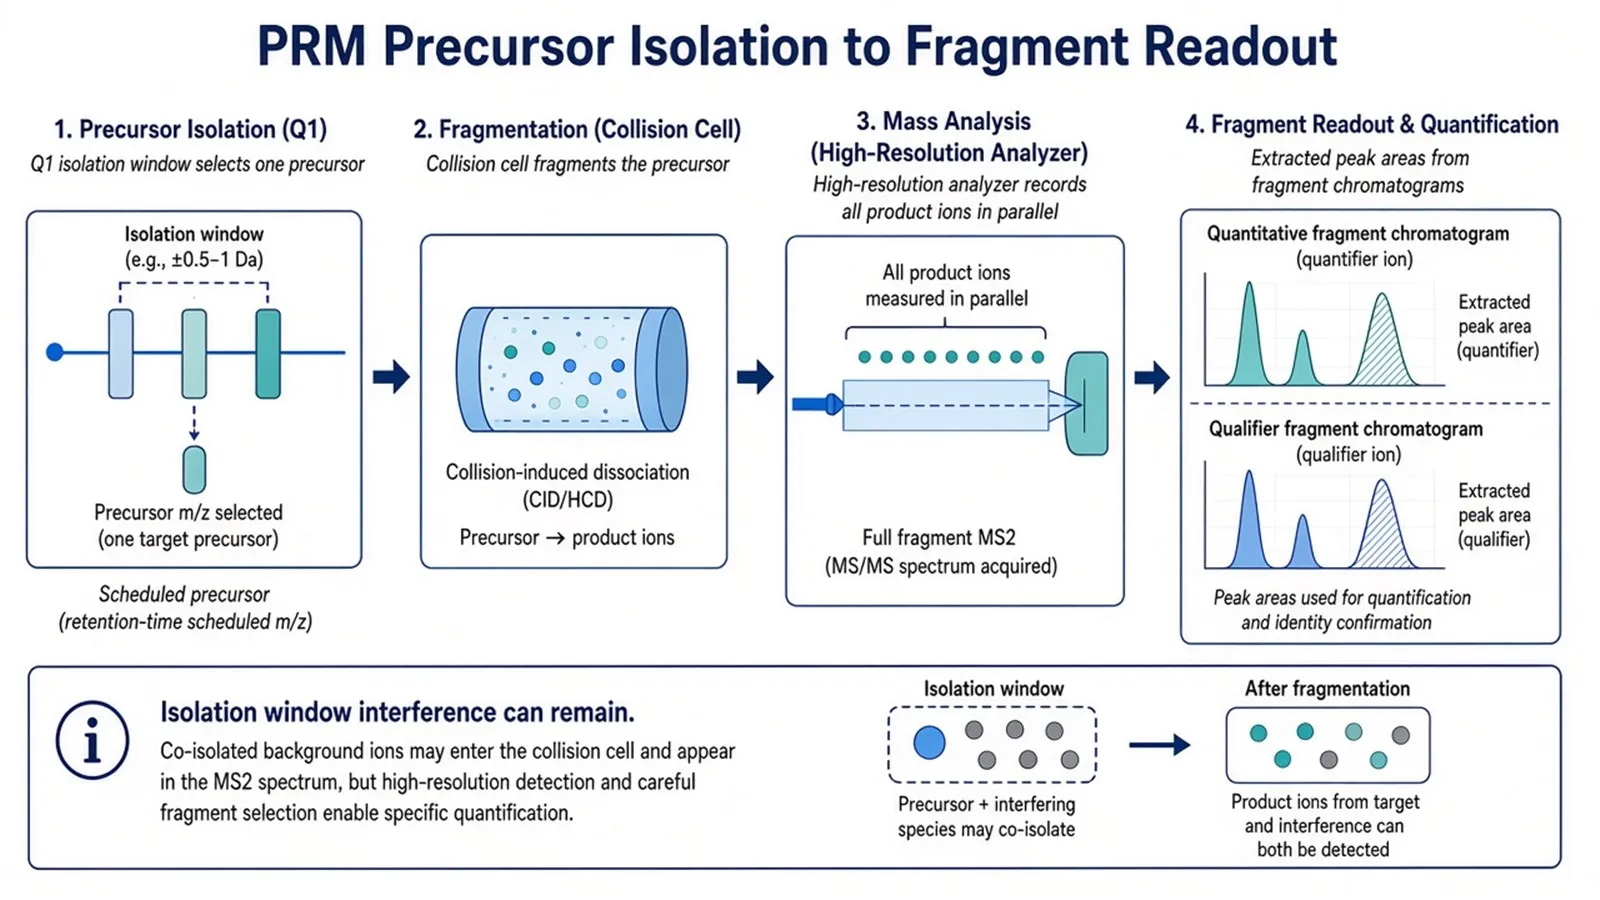
Figure 2. PRM stores the full fragment MS2 of each scheduled precursor; peak areas are extracted after acquisition, while isolation-window interference can still remain.
PRM Versus MRM in Operational Terms
| Operational point | PRM | MRM / SRM |
|---|---|---|
| Instrument class | Quadrupole-Orbitrap | Triple quadrupole |
| What must be set before the run | Precursor list and schedule | Precursor-product transitions |
| Fragment information stored | Full high-resolution MS2 | Selected transitions only |
| Early method development | Often lighter transition picking | Transition optimization is central |
| Multiplex limit | Cycle time set by MS2 fill and scan | High multiplex when transitions are short |
| Typical next use | Validation panel in complex matrices | Mature high-throughput panels |
PRM is often preferred when the panel is still being refined and fragment evidence must be inspected in a complex matrix. MRM remains attractive when a mature transition list must run at very high multiplex on a triple quadrupole. Researchers comparing those trade-offs in more detail can refer to Advantages and Disadvantages of MRM/PRM Techniques.
When PRM Fits
PRM fits when the protein or peptide inventory is already short enough to schedule, the remaining samples can support a targeted assay, and the study needs fragment-level confirmation rather than a new open discovery list.
| PRM often fits | Reassess before scheduling | |
|---|---|---|
| Target status | Closed peptide list from discovery or literature | Open proteome still required |
| Matrix | Complex digests where unit-resolution interference is a concern | Extremely large panels that exceed PRM cycle time |
| Result type | Relative confirmation of nominated peptides | Absolute amount without SIS design |
| Follow-up path | Discovery-to-validation after DDA or DIA |
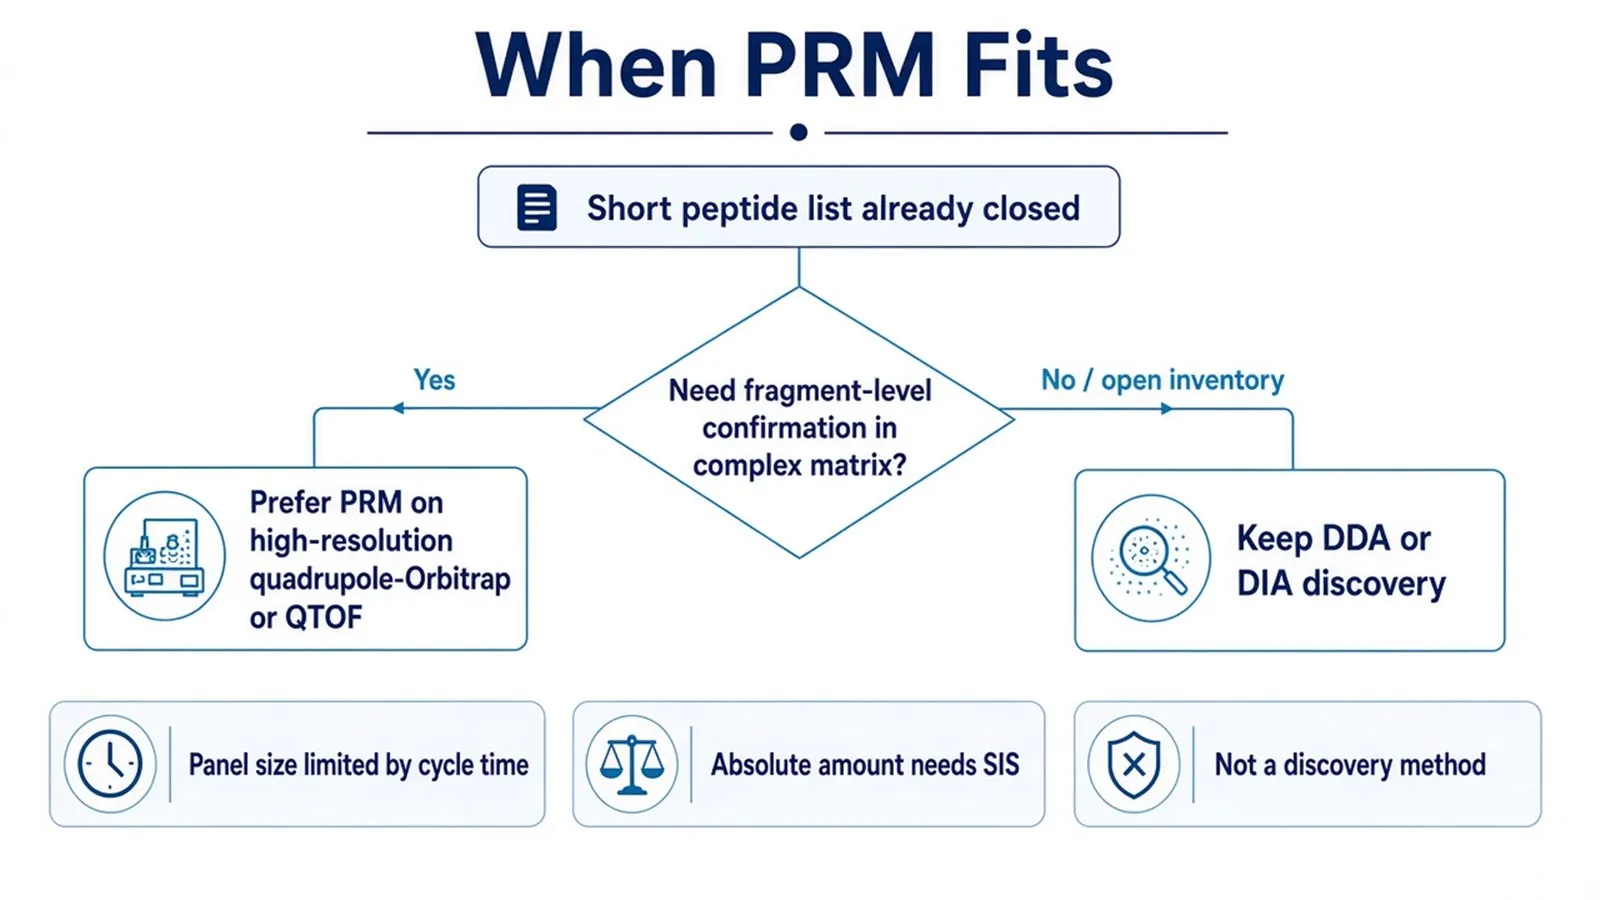
Figure 3. PRM is a validation and targeted quantification route for a closed list; keep DDA or DIA while new protein names are still required.
PRM Limits and Common Uses
1. What Can Limit a PRM Result
Panel size is limited by chromatographic peak width and MS2 cycle time. Shared peptides report gene-family ambiguity, not necessarily a single protein. Digestion and enrichment failures upstream of the SIS spike remain invisible to MS2 resolution. Calling PRM comprehensive because all fragments of a precursor are recorded confuses fragment coverage with proteome coverage.
2. Where PRM Is Commonly Used
PRM is commonly applied to biomarker candidate validation after DDA or DIA, targeted quantification of pathway proteins in plasma or tissue digests, pharmacodynamic time-course panels, and site-aware checks for selected modified peptides when discriminating fragments can be reviewed. Those applications still require declared groups, matched processing, and a clear choice between a relative comparison and an absolute amount.
Once the peptide list, relative-versus-absolute design, and schedule feasibility are defined, the project can be evaluated as a PRM study. Researchers can review the MtoZ Biolabs Parallel Reaction Monitoring (PRM) Service for sample evaluation, peptide-panel feasibility, and workflow planning.
Frequently Asked Questions
1. What is the minimum operational sequence for a PRM run?
Finalize the peptide list, prepare matched digests, schedule precursors, acquire full high-resolution MS2 for those precursors, and extract fragment peak areas. Optional SIS peptides are added when an absolute amount is required.
2. Does PRM replace DDA or DIA discovery?
No. PRM measures only scheduled precursors. Keep DDA or DIA while the protein inventory is still open.
3. Why can PRM method development be faster than MRM?
Because quantitative fragments can be chosen after the full MS2 is recorded. Precursors and retention-time windows must still be scheduled correctly.
4. Does high resolution remove all interference?
No. It improves product-ion discrimination. Co-isolation inside the precursor window and overlapping fragments can remain.
5. When is an SIS required in the workflow?
SIS peptides are commonly used for absolute quantification as part of a calibrated assay with known standard amounts. Relative comparisons of the same peptide can proceed without SIS when sample processing and chromatography remain comparable. SIS peptides can also support peptide identity and retention-time verification.
Reference
-
A.C. Peterson, J.D. Russell, D.J. Bailey, M.S. Westphall, J.J. Coon (2012). Parallel reaction monitoring for high resolution and high mass accuracy quantitative, targeted proteomics. Mol. Cell. Proteomics, 11, 1475-1488. https://doi.org/10.1074/mcp.O112.020131
