



Choose SWATH-MS when relative unlabeled discovery needs sequential-window data-independent acquisition (DIA), library-based fragment extraction, and lower stochastic missingness across many liquid chromatography-tandem mass spectrometry (LC-MS/MS) runs. Choose data-dependent acquisition mass spectrometry (DDA-MS) when real-time top-N precursor selection, cleaner single-precursor MS2 spectra, novel-matrix identification, or a matched DDA spectral library for later SWATH are the priority. SWATH-MS is one sequential-window DIA implementation, not a third class beside DIA and DDA (Gillet et al. 2012). Both routes usually support relative same-protein comparison inside one coherent plan. Neither reports absolute concentration by default, and neither replaces a closed tandem mass tag (TMT) plex or targeted PRM or MRM when those claims fit better.
Researchers who already have defined samples, comparison groups, or a specific project objective can also review the MtoZ Biolabs Deep DIA Proteomics Service and Label-Free Quantitative Proteomics Service for project-specific feasibility and analysis planning.
SWATH-MS vs DDA-MS at a Glance
Dimension |
DDA-MS |
SWATH-MS (DIA implementation) |
|---|---|---|
Precursor selection |
Intensity-triggered top-N MS2 picks |
Consecutive m/z windows fragmented each cycle |
Typical MS2 character |
Often fewer co-fragmented precursors; clearer spectra |
Multiplexed window spectra; higher interference risk |
Common quantitative feature |
MS1 peak-area LFQ (e.g., MaxLFQ) after identification |
Fragment-ion chromatograms extracted with a spectral library |
Missing-value pattern |
More stochastic gaps across runs |
Often more complete matrices; still QC-dependent |
Library role |
Can build libraries for later SWATH or DIA |
Usually library-based extraction; unmatched libraries transfer bias |
Claim type |
Relative discovery or identification-centric work |
Relative unlabeled discovery at cohort scale |
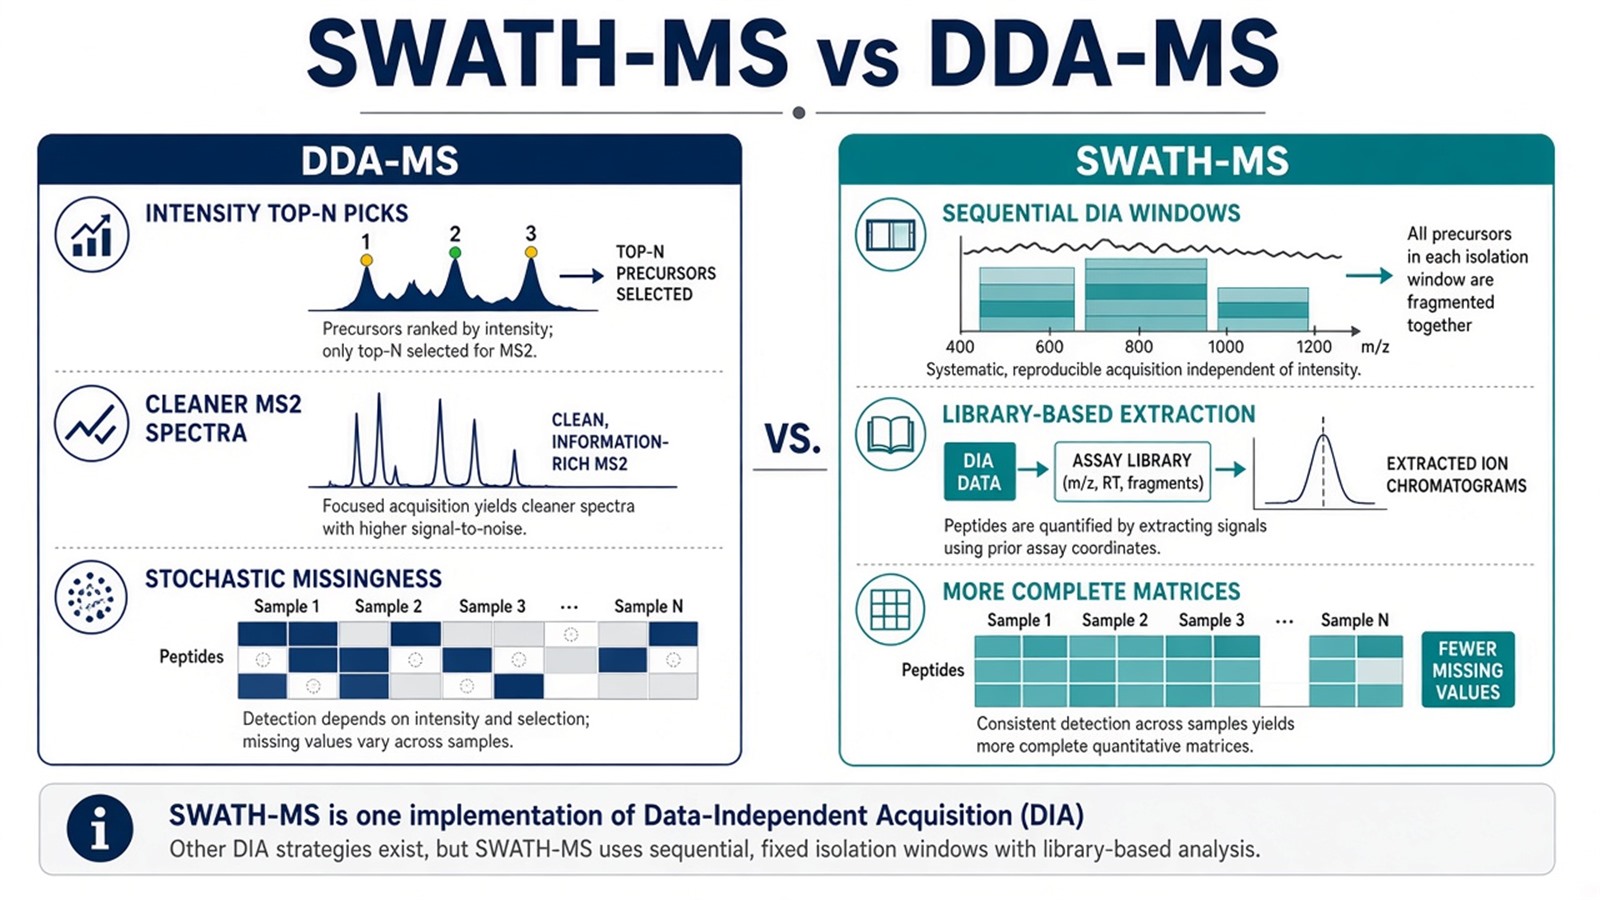
Figure 1. DDA-MS and SWATH-MS differ in precursor selection and quantitative features; SWATH remains inside the DIA acquisition class.
Pick the selection logic that matches the research claim; do not treat SWATH as automatically more accurate for every matrix.
How SWATH and DDA Create Different Research Trade-Offs
DDA-MS: stochastic depth with clearer spectra
DDA-MS surveys MS1, then fragments the most intense precursors. That favors identification confidence and library building, but low-abundance peptides may appear in some runs and not others. Relative quantification in unlabeled DDA often uses aligned MS1 areas after peptide identification when run order and normalization are controlled.
SWATH-MS: sequential DIA windows with library extraction
SWATH-MS divides the precursor range into consecutive isolation windows and fragments all ions in each window on a fixed schedule. Peptide quantities are typically recovered by matching extracted transitions to a spectral library that stores identity, fragments, and retention-time information (Gillet et al. 2012). Matrices are often more complete than stochastic DDA, but chimeric windows, library coverage, and FDR-controlled extraction become part of the method.
Library-free or Direct DIA routes can reduce dependence on a separate empirical library while still using scheduled-window acquisition. They change extraction strategy inside DIA; they do not turn SWATH into a separate omics category.
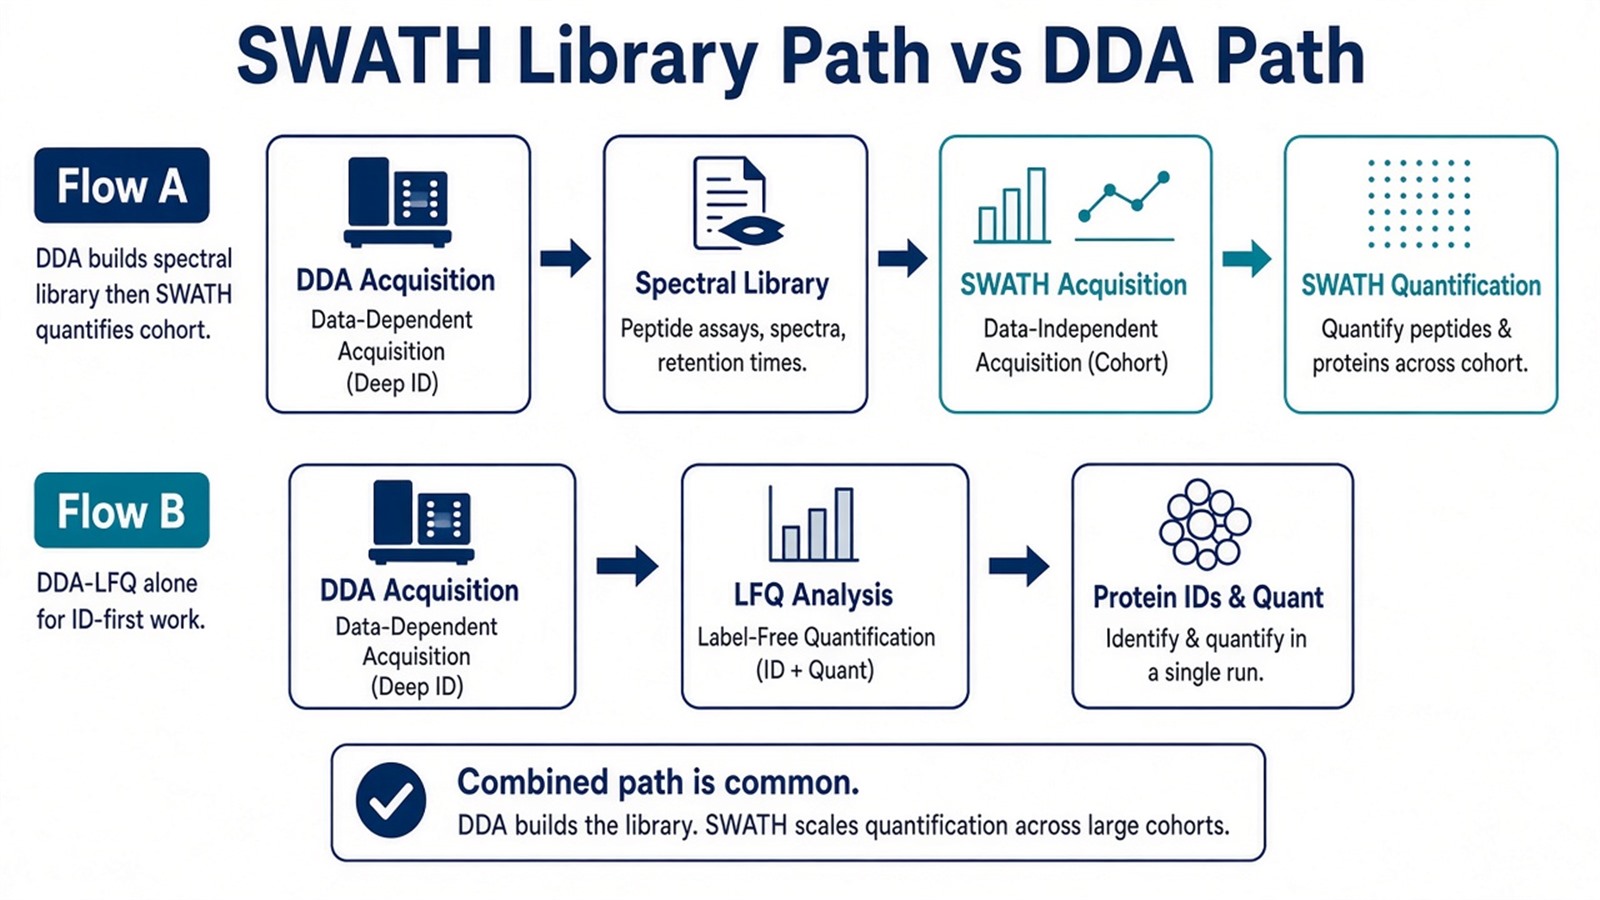
Figure 2. A frequent suitable design is DDA library construction on matched material, then SWATH-MS cohort quantification; DDA alone still fits identification-first work.
For definitions of DIA versus SWATH beyond this selection page, see What Is DIA and SWATH? Understanding Data-Independent Acquisition in Proteomics? and What Is SWATH Mass Spectrometry. For broader DIA versus DDA selection, see DIA vs DDA Proteomics: Key Differences and Applications Explained. For LFQ-focused stacks, see DIA vs. DDA in Label-Free Quantitative Proteomics: A Comparative Analysis.
Judgment Criteria: Which Technique Fits Your Research
Study need |
Prefer SWATH-MS / SWATH-style DIA |
Prefer DDA-MS |
Reassess both |
|---|---|---|---|
Cohort quantification |
Lower stochastic missingness across many runs |
Identification-first pilot or library construction |
Absolute concentration required |
Spectral priority |
Fragment chromatogram completeness with library extraction |
Cleaner single-precursor MS2 for confident ID |
Closed peptide panel ready for PRM or MRM |
Library logistics |
Matched empirical library ready |
Build a matched DDA library for later SWATH |
Unmatched library with no library-free DIA path |
Sample complexity |
Complex digests where consistent coverage matters |
Interactome or IP samples needing clearer MS2 |
Chemistry forces a closed TMT plex |
Analysis capacity |
DIA or SWATH software and FDR QC in place |
MaxQuant-centered DDA-LFQ plan in place |
No stable chromatography or batch plan |
Prefer SWATH-MS when relative unlabeled discovery needs scheduled-window completeness and a workable library or library-free DIA path. Prefer DDA-MS when identification clarity, MaxLFQ-style unlabeled files, or library building comes first. Prefer a combined path when library quality and cohort throughput must both be planned. Prefer Direct DIA when empirical library construction is the bottleneck. Prefer PRM or MRM when targets are already closed.
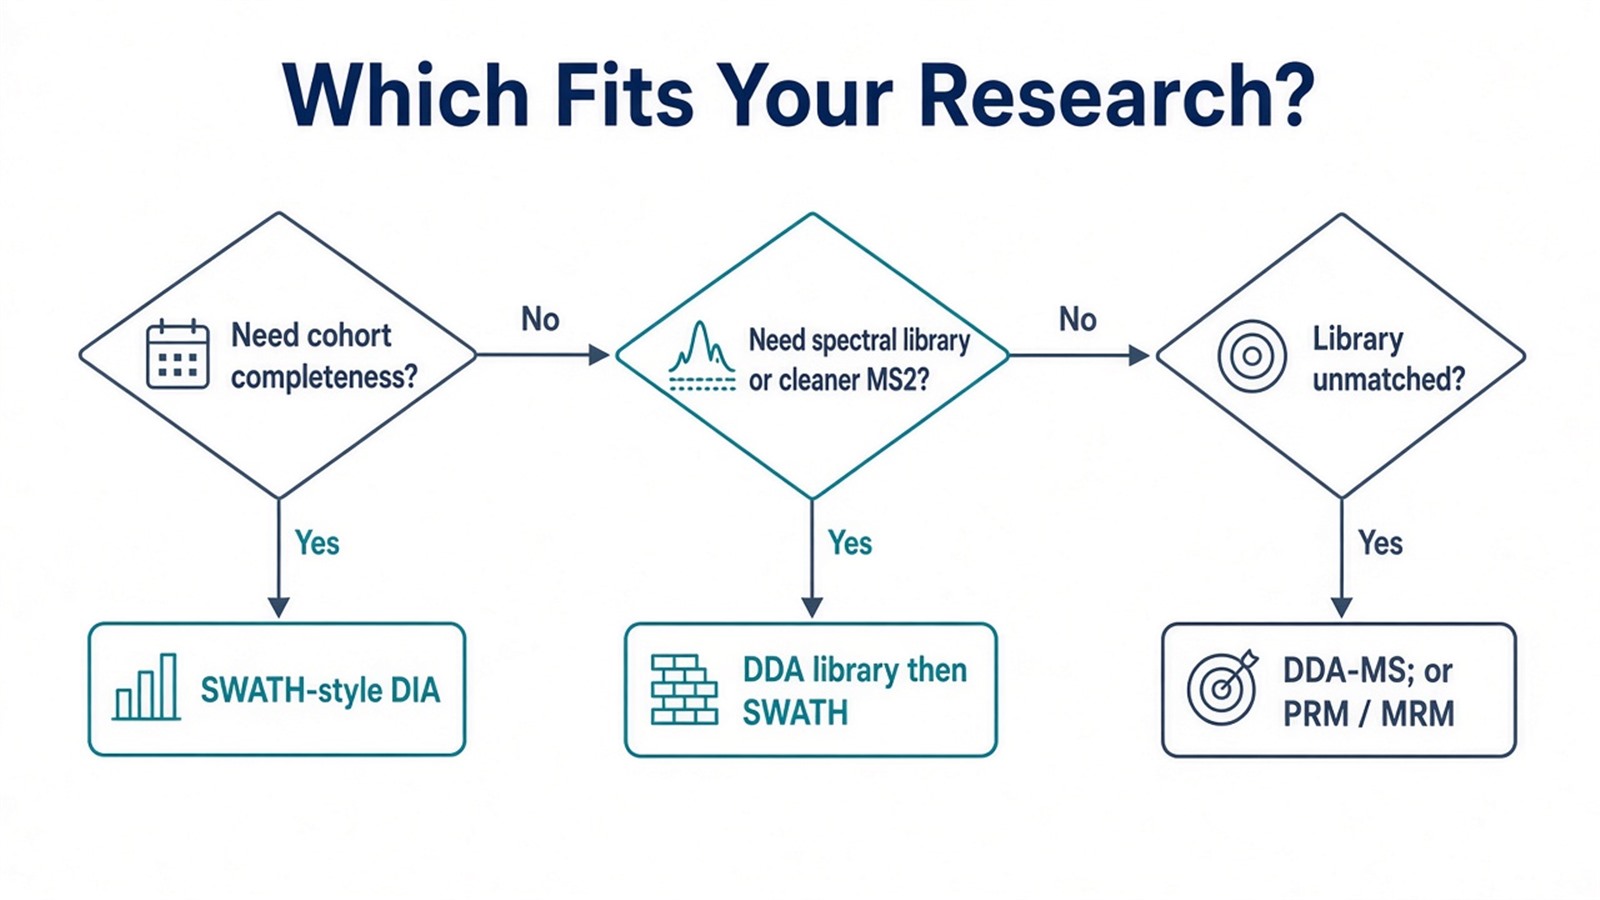
Figure 3. Suitability starts from missing-value tolerance, spectral needs, and library logistics, not from naming SWATH as universally better.
For DIA advantages and limits, see Advantages and Disadvantages of DIA in Protein Quantification and Strengths and Limitations of DIA in Protein Quantification. For library-free options, see Direct DIA: High-Efficiency Proteomics without a Spectral Library. For analysis tooling, see Essential Analysis Tools for Label-Free Proteomics: A Comprehensive Review of MaxQuant and DIA-NN.
Limits and Application Boundaries
SWATH-MS does not remove window interference. DDA-MS does not remove stochastic missingness. Calling either universally more reproducible or more sensitive overstates method class alone. Ordinary intensities from both routes remain relative unless matched standards and calibration are added. Unmatched libraries transfer tissue or batch bias into SWATH matrices. Cross-batch comparability needs design and QC, not only the SWATH name.
Typical applications:
- DDA-MS: novel-matrix identification, interactome or immunoprecipitation studies needing clearer MS2, DDA-LFQ discovery, and spectral-library construction for later SWATH.
- SWATH-MS: treatment or genotype screens, larger unlabeled cohorts, complex-matrix relative discovery, and projects that may re-extract stored files under updated libraries.
- Combined path: DDA library on matched material, then SWATH quantification across the cohort, or SWATH discovery followed by PRM confirmation of a shortlist.
Diagnostic cutoffs sit outside ordinary research SWATH or DDA reporting unless a separately validated assay framework is in place.
Once the acquisition choice, library or library-free route, and relative comparison design are defined, the project can be evaluated as a SWATH-style DIA or DDA study. Researchers can review the MtoZ Biolabs Deep DIA Proteomics Service for SWATH-style DIA planning and the Label-Free Quantitative Proteomics Service for unlabeled DDA options. For acquisition plus TMT stacks, see DDA vs. DIA vs. TMT in LC-MS/MS Proteomics: How to Choose the Right Strategy.
Frequently Asked Questions
1. Is SWATH-MS different from DIA?
SWATH-MS is one sequential-window DIA implementation, commonly with library extraction. It is not a third class beside DIA and DDA.
2. Is SWATH always better than DDA for quantification?
No. SWATH often improves cohort completeness for relative unlabeled discovery. DDA can still fit identification-first work, MaxLFQ plans, and library building when QC is controlled.
3. Do I need a spectral library for SWATH?
Classic SWATH workflows are library-based. Library-free DIA can reduce that bottleneck but still needs FDR control and interference review.
4. Can DDA and SWATH be used in the same project?
Yes. A common sequence is DDA for library construction, then SWATH for cohort quantification, or SWATH discovery followed by PRM confirmation.
5. Do SWATH-MS and DDA-MS provide absolute protein concentration?
Not by default. Both report relative values inside a coherent plan unless matched standards and calibration are added.
Reference
- L.C. Gillet, P. Navarro, S. Tate, H. Rost, N. Selevsek, L. Reiter, R. Bonner, R. Aebersold (2012). Targeted data extraction of the MS/MS spectra generated by data-independent acquisition: a new concept for consistent and accurate proteome analysis. Mol. Cell. Proteomics, 11, O111.016717. https://doi.org/10.1074/mcp.O111.016717
- J. Cox, M.Y. Hein, C.A. Luber, et al. (2014). Accurate proteome-wide label-free quantification by delayed normalization and maximal peptide ratio extraction, termed MaxLFQ. Mol. Cell. Proteomics, 13, 2513-2526. https://doi.org/10.1074/mcp.M113.031591
