



Sequential window acquisition of all theoretical fragment ion spectra (SWATH-MS) is often applied in plant proteomics when unlabeled digests from leaves, roots, seeds, or other tissues must be compared across genotypes, treatments, or time points on separate liquid chromatography-tandem mass spectrometry (LC-MS/MS) injections (Gillet et al. 2012). Label-free chemistry and a windowed data-independent acquisition (DIA) schedule support open relative contrasts in cohorts that may still grow. The guide below maps SWATH application choices for plant projects from matched prep through extraction. It is not a crop breeding promise, not a pathway-enrichment manual, and not a generic SWATH theory page.
Plant matrices add prep constraints that animal cell lines often avoid. Cell walls, polysaccharides, phenolics, and pigments can suppress digestion and chromatography when cleanup is unequal across groups (Wisniewski et al. 2009). SWATH does not remove those compounds. Matched extraction and digestion across treatments remain the application gate before windowed acquisition starts.
Plant Application Contexts Where SWATH Fits
Three plant application patterns appear most often.
Stress or treatment contrasts on the same tissue type. Drought, salt, temperature, or hormone treatments compared to control plants need matched harvest timing, tissue dissection, and prep batches interleaved with acquisition order.
Genotype or accession panels with open enrollment. Separate unlabeled injections fit when new lines or harvest dates may still arrive.
Tissue comparison within one genetic background. Root versus leaf versus flower contrasts need explicit tissue prep rules so abundance differences are not confounded with extraction chemistry.
SWATH plant applications still require a predefined relative contrast on matched digests, not a proteome survey keyword alone.
Researchers who need general label-free SWATH pretreatment rules can refer to SWATH-MS Label-Free Protein Quantification Technology Analysis.
Study Designs for Plant SWATH Cohorts
Application quality follows plant-specific design choices.
Tissue and developmental stage must be fixed in the contrast map. Mixing seedlings and mature leaves in one arm without declaration creates false regulation lists.
Matched extraction chemistry. Phenol-assisted or trichloroacetic acid and acetone precipitation routes are common when secondary metabolites interfere. The same route must apply to every arm in the contrast.
Database and library provenance. Model species with solid reference proteomes fit library or library-free extraction when species and digestion match the search space (Demichev et al. 2020). Non-model species need an explicit database or library plan. Missing annotations limit functional wording but do not remove the need for quantitative QC on the matrix.
Enrollment and batch plan. Greenhouse batch, harvest day, and extractor version should not track treatment unless that tracking is part of the design.
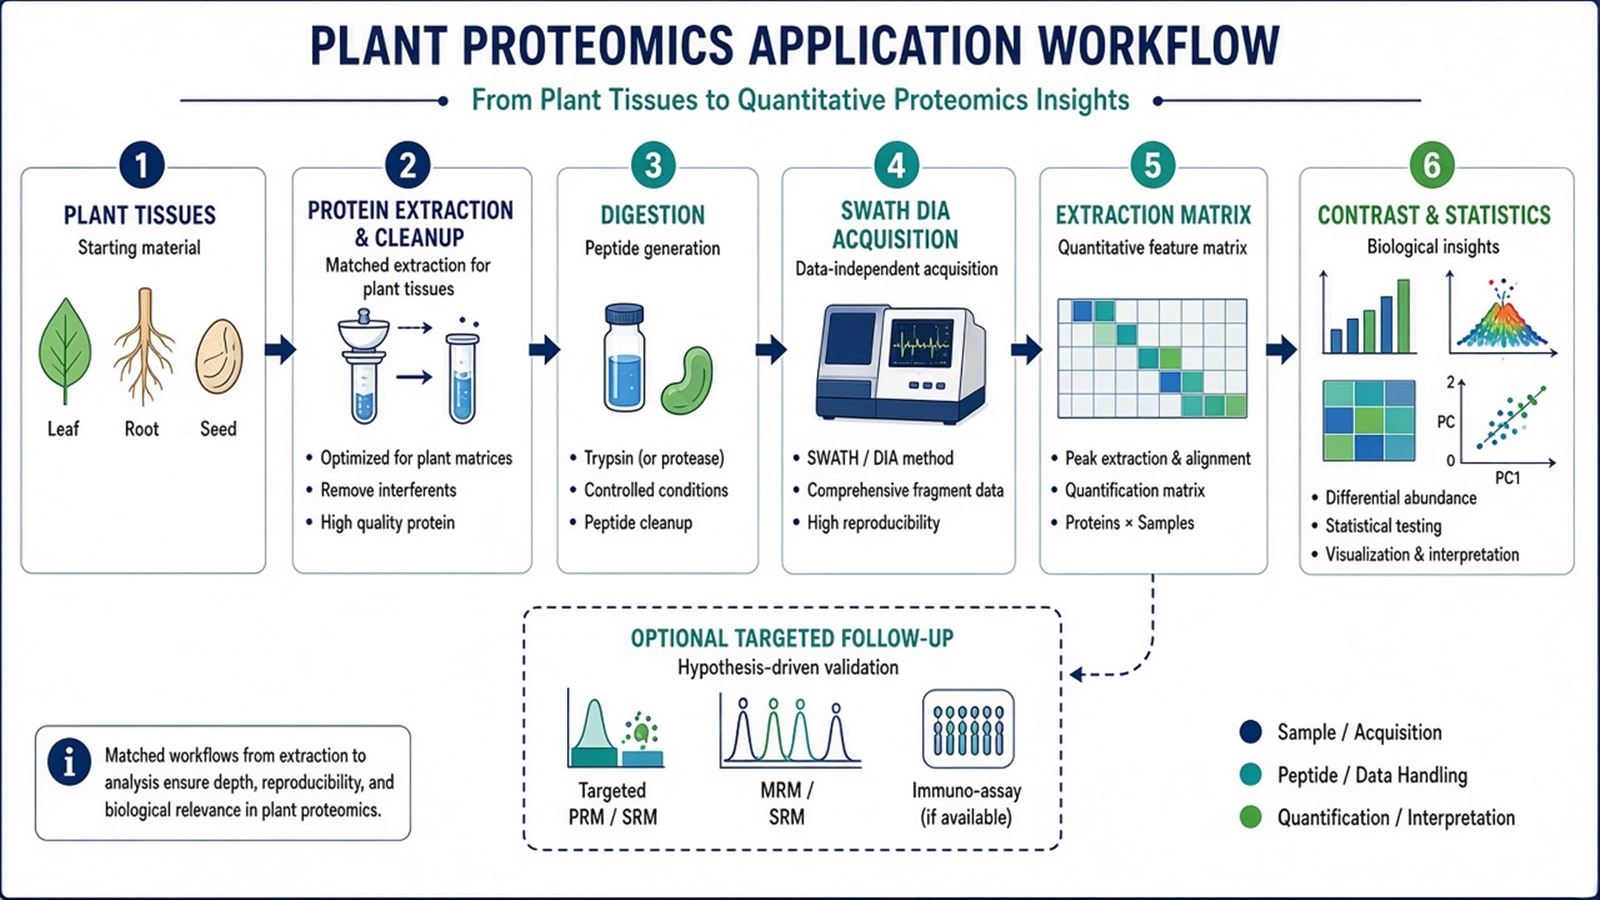
Figure 1. Plant SWATH applications depend on matched tissue prep and a declared contrast before windowed acquisition and extraction.
High-throughput cohort framing for discovery lists is covered in High-throughput SWATH-MS Quantitative Proteomics for Biomarker Discovery at the generic cohort level. This page stays on plant matrix and contrast choices.
Shared Workflow From Plant Sample to SWATH Matrix
A practical plant SWATH spine usually runs as follows.
- Confirm species, tissue, contrast, and relative claim.
- Harvest with matched timing; record batch and storage.
- Extract proteins with one declared plant-compatible route; remove debris and interfering metabolites.
- Digest, clean peptides, and desalt to LC-ready loads shared across arms.
- Run pilot injections when the tissue or prep route is new.
- Acquire SWATH windows under a locked plan across the cohort.
- Extract peptides under FDR and interference rules; test the predefined contrast only (Rosenberger et al. 2017).
- Optional pathway overlay or targeted remeasurement on a short list after QC.
Steps 2 through 5 are where plant applications succeed or fail. Step 7 belongs with DIA Data Analysis Methods in Proteomics. Skipping pilot prep on high-pigment tissue often forces repeat extractions after the full SWATH batch is scheduled.
Missing values in plant digests often track low abundance and prep loss, not only acquisition chance (Lazar et al. 2016). Enrichment cannot repair treatment-confounded harvest batches.
Parameters and Limits in Plant SWATH Applications
Parameter |
Why it matters in plant work |
Failure mode if ignored |
|---|---|---|
Tissue type and stage |
Sets extraction and contrast |
Mixed organs read as regulation |
Metabolite-aware cleanup |
Protects digestion and LC |
Suppressed or missing peptides by arm |
Matched input (fresh weight or total protein) |
Stabilizes yield |
Load bias mimics biology |
Species database or library plan |
Bounds identification |
Wrong species search space |
Window scheme locked to cohort |
Keeps extraction valid |
Mid-study window changes |
Run-order interleaving |
Separates batch from treatment |
Greenhouse day tracks group |
Follow-up on short lists |
Separates discovery from confirmation |
Pathway arrows as mechanism proof |
Limits follow the same table. SWATH does not ensure every low-abundance regulator in every tissue. Retrospective reanalysis still needs a declared extractor and contrast. Breeding markers, yield predictions, and trait claims need genetics and validation outside an open SWATH table. Multi-omic overlay can suggest hypotheses. It does not make protein ratios interchangeable with transcript units.
TMT may fit a freezable multi-arm plant set that can be labeled the same day. Open plant nurseries with staggered harvests more often stay on unlabeled SWATH separate injections.
Researchers who need three-axis route selection can refer to Protein Quantification Method. For general matched prep spine detail, see Quantitative Protein Analysis.
How to Choose SWATH for the Next Plant Project
Use matrix, enrollment, and follow-up goal together.
Project need |
SWATH application plan that often fits |
Recheck before committing |
|---|---|---|
Treatment versus control, same tissue |
Matched plant prep, SWATH, locked extractor |
Harvest day equals treatment |
Many genotypes arriving over time |
Unlabeled SWATH separate injections |
Forced TMT without freezable set |
Root versus shoot within one line |
Paired tissue prep rules per organ |
Different prep routes per organ unlabeled |
High-pigment tissue |
Metabolite-aware cleanup pilot |
Full cohort without pilot LC |
Non-model species |
Declared database or library plan |
Functional claims beyond ID support |
Short list after discovery |
Same prep, targeted PRM panel |
Open SWATH p-values as final decision |
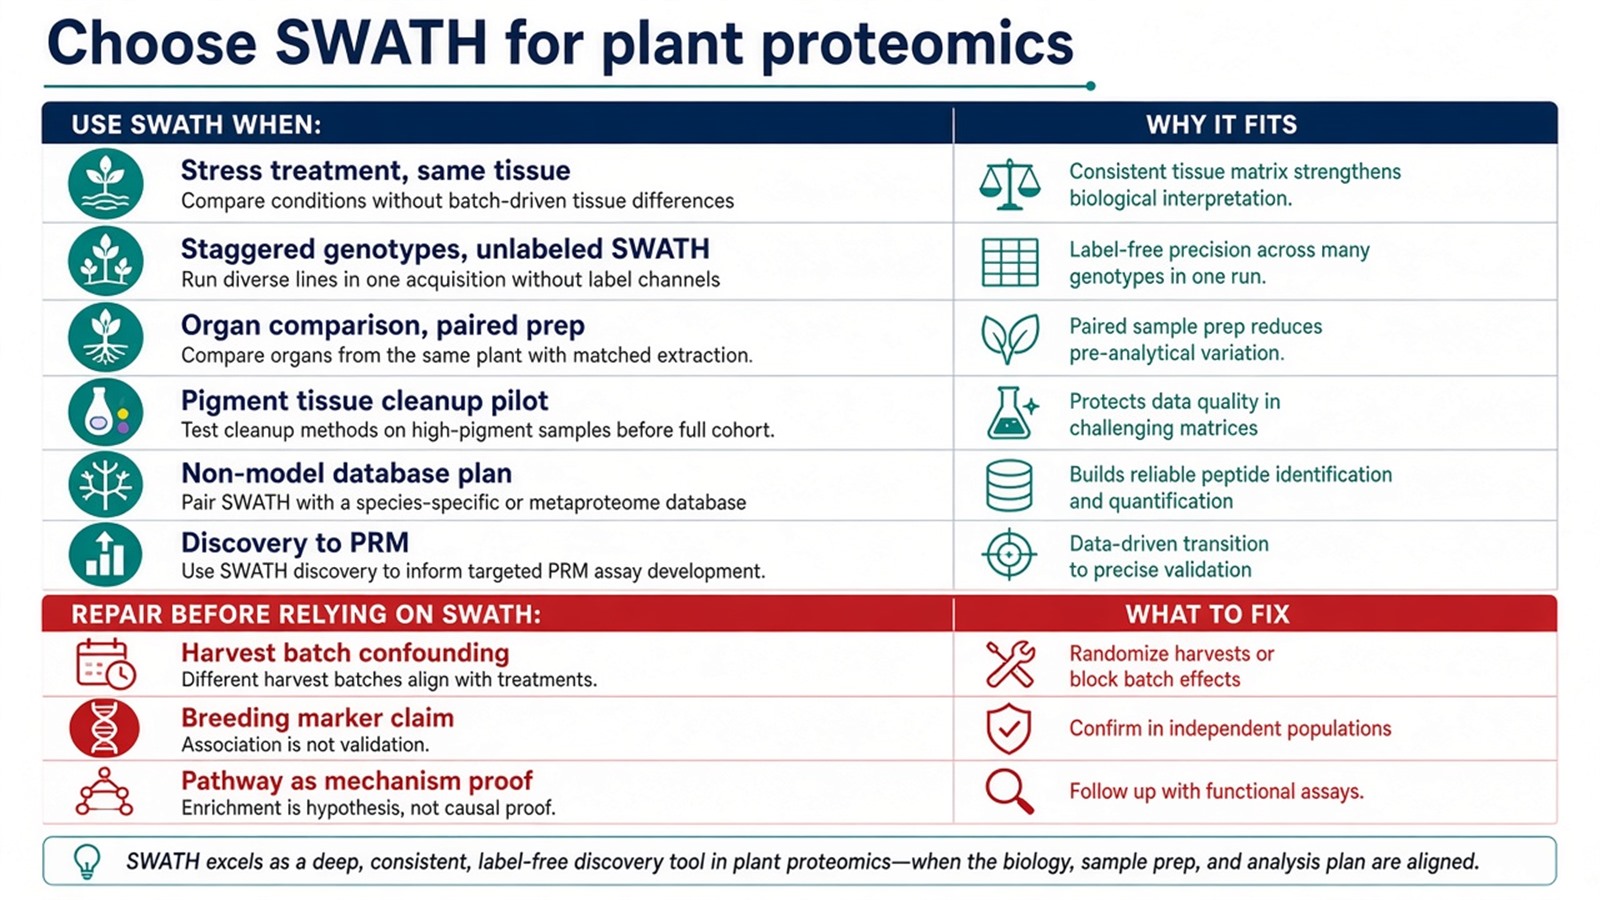
Figure 2. Choose plant SWATH when matched tissue prep and a relative contrast fit; pigment prep, batch confounding, and breeding claims need separate plans.
Unlabeled SWATH more often fits open plant cohorts and staggered harvests. TMT more often fits a closed set harvested for one labeling day. Targeted confirmation more often fits when trait or mechanism decisions depend on remeasuring a short peptide list. Exceptions exist when only a pilot tissue comparison is planned before scaling prep.
Teams that already have species, tissue map, contrast, and follow-up goal can also review the MtoZ Biolabs Label-Free Quantitative Proteomics Service for project-specific plant SWATH planning.
Frequently Asked Questions
1. Is SWATH required for all plant proteomics projects?
No. SWATH fits open relative contrasts on unlabeled separate injections. Small closed target lists may move to targeted assays after names freeze.
2. Can one SWATH run replace replicated plant harvests?
No. Biological replicates and matched prep remain required. Acquisition mode does not replace replicate design.
3. Does a weak reference genome block SWATH quantification?
It constrains identification and annotation. Quantitative extraction still needs a declared search space and QC on the matrix.
4. When should a custom ion library be planned?
When library-based extraction is chosen and public libraries do not match species, digestion, and chromatography. Library-free routes are an alternative with their own QC rules.
5. Can pathway enrichment prove stress mechanism from one SWATH contrast?
No. Enrichment interprets a contrast-tested list. Mechanism claims need orthogonal evidence and often targeted follow-up.
MtoZ Biolabs can support plant SWATH application planning when species, tissue type, contrast design, and the intended relative claim are available.
If further evaluation of metabolite-aware prep, species database strategy, window locking, or discovery-to-targeted staging is needed, project details can be submitted for a feasibility review.
Reference
- V. Demichev, C. Messner, S. Vernardis, K.S. Lilley, M. Ralser (2020). DIA-NN: neural networks and interference correction enable deep proteome coverage in high throughput. Nat. Methods, 17, 41-44. https://doi.org/10.1038/s41592-019-0638-x
- L.C. Gillet, P. Navarro, S. Tate, H. Rost, N. Selevsek, L. Reiter, R. Bonner, R. Aebersold (2012). Targeted data extraction of the MS/MS spectra generated by data-independent acquisition: a new concept for consistent and accurate proteome analysis. Mol. Cell. Proteomics, 11, O111.016717. https://doi.org/10.1074/mcp.O111.016717
- C. Lazar, L. Gatto, M. Ferro, C. Bruley, T. Burger (2016). Accounting for the multiple natures of missing values in label-free quantitative proteomics data sets to compare imputation strategies. J. Proteome Res., 15, 1116-1125. https://doi.org/10.1021/acs.jproteome.5b00981
- G. Rosenberger, S. Liu, J. Röst, H.L. Ludwig, C. Buil, B. Borchers, R. Aebersold, L. Collins (2017). Statistical control of peptide and protein error rates in large-scale targeted data-independent acquisition analyses. Nat. Commun., 8, 14224. https://doi.org/10.1038/ncomms14224
- J.R. Wisniewski, A. Zougman, N. Nagaraj, M. Mann (2009). Universal sample preparation method for proteome analysis. Nat. Methods, 6, 359-362. https://doi.org/10.1038/nmeth.1322
