



Protein quantification technology with TMT labeling quantitation is a multiplex relative workflow for discovery-scale proteomes. Tandem mass tags (TMT) attach channel-specific isobaric labels to digests so peptides from many samples share one precursor mass on liquid chromatography-tandem mass spectrometry (LC-MS/MS), then release reporter ions after fragmentation for within-plex ratios (Thompson et al. 2003). In project language, TMT answers predefined group contrasts inside a labeled set. It does not report mass or molar concentration unless a matched stable isotope-labeled internal standard (SIS) is added for that peptide, and it does not replace closed-panel multiple reaction monitoring (MRM) or parallel reaction monitoring (PRM) verification. The guide below maps TMT scope, workflow, locked parameters, limits, and project fit versus label-free quantification (LFQ) and targeted routes. It is not a multi-omics brochure, not a clinical biomarker catalog, and not a claim that mixing alone guarantees accuracy.
Ratio compression from co-isolated precursors remains a known MS2 TMT limit; MS3 or synchronous precursor selection MS3 (SPS-MS3) can reduce that mixing on compatible instruments (Ting et al. 2011; McAlister et al. 2014). Unmatched protein load or digest still breaks reporter biology after normalization (Bantscheff et al. 2012).
In MtoZ Biolabs quantitative offerings, TMT labeling quantitation is scoped after plex size, channel map, matrix, relative claim, fractionation plan, and quality control (QC) design are written in the brief. Service intake appears in TMT Quantitative Proteomics Analysis Service.
What TMT Labeling Quantitation Must Cover
Four layers belong in the brief and stay separated.
Contrast layer freezes biological groups, replicates, and the relative claim inside the plex. Open “find anything interesting” without a contrast is outside a locked TMT record.
Plex-and-channel layer names kit size, which samples occupy which tags, bridge or reference channels if more than one plex is planned, and randomization rules. Kit channel count is not a biological result.
Sample-and-prep layer names matrix, matched protein input, digest enzyme, labeling completeness checks, quench, and cleanup. Residual amine buffers compete for reagent and leave unlabeled peptides.
Acquisition-and-report layer names MS2 versus MS3 reporter strategy, fractionation if used, search and reporter software, and whether a later targeted verify step is planned.
TMT is isobaric at MS1; a reporter ratio means biology only after matched load, complete labeling, and a declared channel map.
Isobaric chemistry and naming detail appear in What Is TMT Isobaric Labeling?. This page stays on comprehensive project framing for TMT protein quantification.
How TMT Compares with LFQ and Targeted MRM/PRM
TMT, LFQ, and targeted monitoring answer different design objects.
TMT mixes labeled channels into one LC-MS/MS path, which removes run-to-run retention drift within that plex and supports many samples under one chromatographic method. LFQ keeps each digest on its own injection and grows the file list without chemical tags. When each sample must stay unlabeled or the cohort keeps expanding, Label-Free Quantitative Proteomics Service framing fits better than forcing a plex.
After a discovery TMT list freezes, MRM or PRM can remeasure named peptides for closed-panel confirmation. Route contrast across MRM, LFQ, TMT, and PRM appears in Comparison of Mass Spectrometric MRM with Other Quantitative Methods in Proteomics. TMT discovery ratios are not automatic absolute amounts and are not antibody epitope assays.
End-to-End Workflow for TMT Labeling Quantitation
A practical comprehensive TMT spine usually runs as follows.
- Freeze contrast, biological n, and relative claim for the protein quantification question.
- Choose plex size and draft the channel map, including bridge channels if multiple plexes are required.
- Match protein input, digest, and cleanup across channels; verify labeling completeness on pilot aliquots.
- Label, quench, and mix; keep residual free reagent out of the low-mass reporter region.
- Fractionate when depth is required; declare the fractionation recipe for every plex.
- Acquire MS2 or MS3 reporter methods with isolation and collision settings locked from the pilot.
- Search, extract reporters, normalize, and test only the predefined contrast.
- Review ratio-compression flags, missing channels, and QC bridge behavior before reporting.
- Hand named targets to MRM/PRM only when a closed verification claim is declared.
Steps 1 through 4 protect the labeled sample record. Steps 5 through 9 produce the quantitative TMT record.
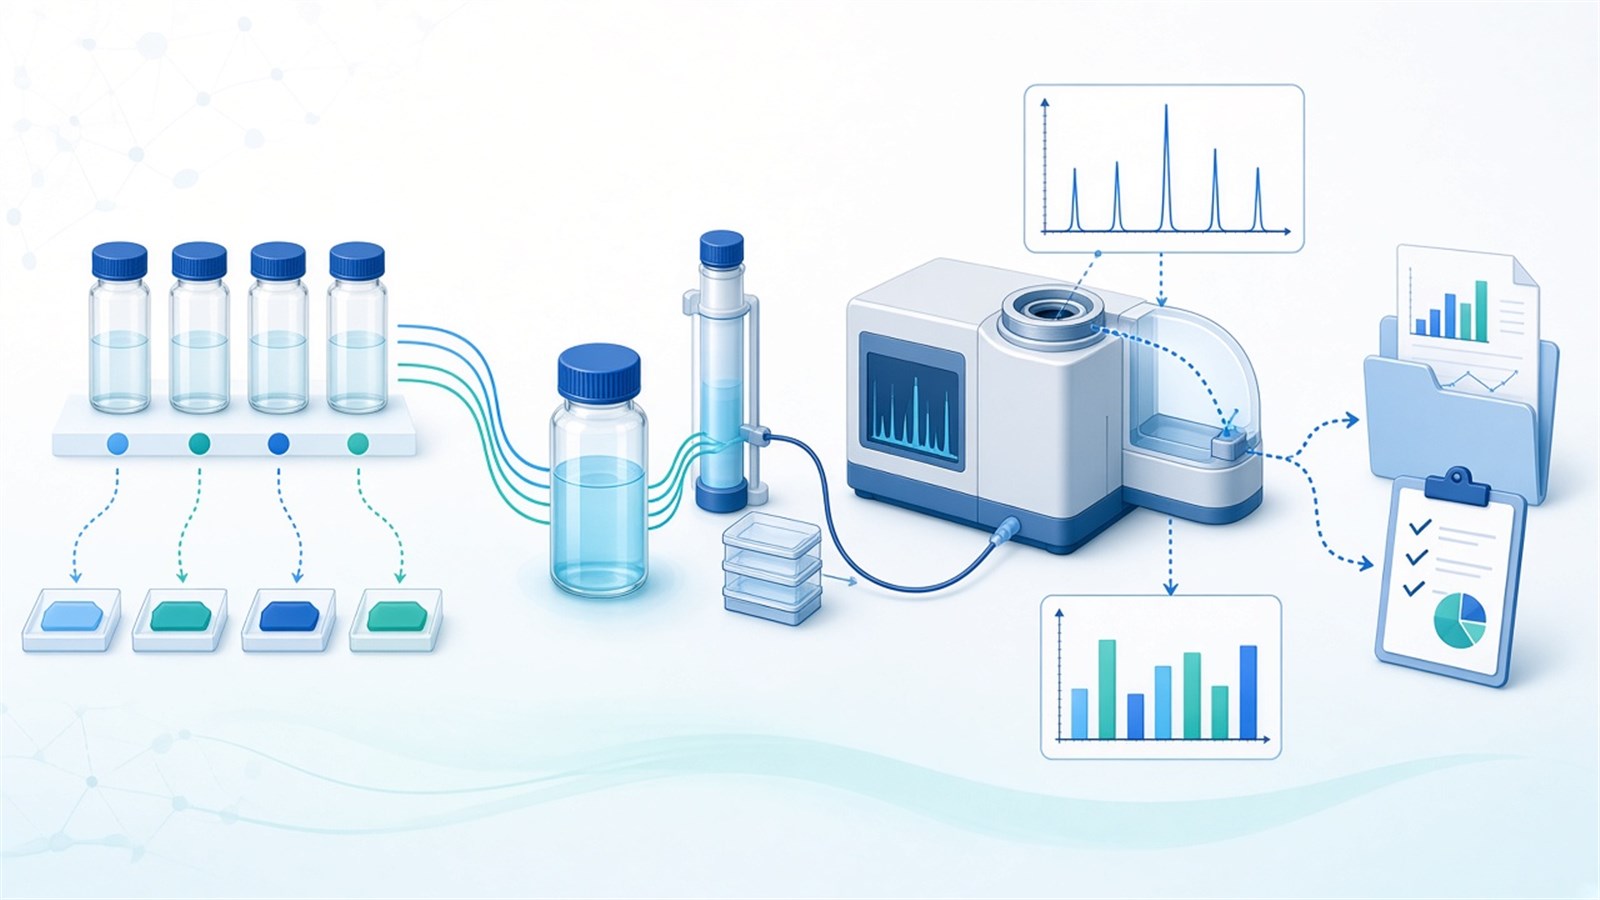
Figure 1. TMT protein quantification depends on a locked channel map and matched prep before reporter ratios are tested as biology.
Ratio-compression mitigation detail appears in TMT MS3 and SPS-MS3: How to Mitigate Ratio Compression and Enhance Quantitative Accuracy.
Sample and Method Parameters That Must Be Locked
Layer |
Parameter that must be declared |
Failure mode if ignored |
|---|---|---|
Contrast |
Groups, replicates, relative claim |
Post-hoc story hunting |
Plex |
Kit size and channel map version |
Channel bias as biology |
Bridge |
Reference design across plexes |
Multi-plex batches not comparable |
Load |
Matched protein input per channel |
Systematic channel offset |
Digest |
Enzyme and missed-cleavage policy |
Unequal amines for labeling |
Labeling |
Completeness check and quench |
Split tagged/untagged forms |
Isolation |
MS2 window or MS3 reporter strategy |
Ratio compression |
Fractionation |
Recipe matched across plexes |
Depth bias between batches |
QC |
Pools, blanks, bridge behavior |
Undetected labeling or instrument drift |
Deliverable |
Channel map, method, search settings |
Report rebuilt after acquisition |
Limits follow the same table. Multiplex does not remove the need for biological replicates. Mixing before LC-MS/MS reduces technical run variance inside a plex; it does not fix unmatched harvest or incomplete labeling. Clinical diagnosis, drug-target proof, and multi-omics network language need separate study design beyond the TMT table.
How to Choose TMT Labeling Quantitation
Use plex need, list status, claim unit, and follow-up intent together.
Project need |
Route that often fits |
Recheck before committing |
|---|---|---|
Many groups, one mixed LC path, relative discovery |
TMT labeling quantitation |
Force LFQ when tags are unnecessary |
Expanding unlabeled cohort |
LFQ or DIA |
Fit every new sample into one TMT kit |
Closed peptide verification after discovery |
MRM or PRM follow-up |
Re-run TMT as a targeted assay |
Strong ratio-compression risk on MS2 |
MS3 / SPS-MS3 TMT plan |
Ignore isolation purity |
Amount on named peptides |
Targeted SIS methods |
Reporter height as concentration |
Single small contrast, no multiplex need |
LFQ may be simpler |
Buy plex chemistry without channel pressure |
Incomplete labeling on pilot |
Fix prep and amines first |
Full cohort on failing label efficiency |
Figure 2. Choose TMT when multiplex relative discovery fits; keep LFQ for unlabeled growth and MRM/PRM when a closed verify claim appears.
Teams with channel maps, matched material plans, and QC-inclusive plex designs can review the same MtoZ Biolabs TMT service entry when those fields are declared in the project record.
Frequently Asked Questions
1. Does TMT always beat LFQ on accuracy because samples are mixed?
Mixing reduces run-to-run technical variance inside a plex. It does not repair unmatched load, incomplete labeling, or MS2 ratio compression.
2. Are TMT reporter ratios absolute protein amounts?
No. Default TMT reports relative channel ratios. Absolute units need peptide-level SIS designs outside standard discovery TMT.
3. How many samples can one TMT experiment hold?
Commercial kits set channel count. Study design decides which biological samples and bridges occupy those channels, including multi-plex layouts.
4. When should projects add MS3 or SPS-MS3?
When MS2 pilots show clear ratio compression or co-isolation risk and the instrument class supports MS3 reporter acquisition.
5. Should every TMT discovery study continue into MRM or PRM?
Only when a closed peptide list and verification claim are declared. Discovery ratios alone are not a targeted assay.
MtoZ Biolabs can discuss TMT labeling quantitation scoping when plex design, matrix, and contrast are documented.
If further evaluation of channel maps, MS3 options, or LFQ versus TMT fit is needed, project details can be submitted for a feasibility review.
Reference
- A. Thompson, J. Schäfer, K. Kuhn, S. Kienle, J. Schwarz, G. Schmidt, T. Neumann, R. Johnstone, A.K. Mohammed, C. Hamon (2003). Tandem mass tags: a novel quantification strategy for comparative analysis of complex protein mixtures by MS/MS. Anal. Chem., 75, 1895-1904. https://doi.org/10.1021/ac0262560
- L. Ting, R. Rad, S.P. Gygi, W. Haas (2011). MS3 eliminates ratio distortion in isobaric multiplexed quantitative proteomics. Nat. Methods, 8, 937-940. https://doi.org/10.1038/nmeth.1714
- G.C. McAlister, D.P. Nusinow, M.P. Jedrychowski, M. Wühr, E.L. Huttlin, B.K. Erickson, R. Rad, W. Haas, S.P. Gygi (2014). MultiNotch MS3 enables accurate, sensitive, and multiplexed detection of differential expression across cancer cell line proteomes. Anal. Chem., 86, 7150-7158. https://doi.org/10.1021/ac502040v
- M. Bantscheff, S. Lemeer, H.U. Haab, B. Kuster (2012). Quantitative mass spectrometry in proteomics: a critical review. Anal. Bioanal. Chem., 404, 939-965. https://doi.org/10.1007/s00216-012-5892-4
