Parallel reaction monitoring (PRM) is a targeted quantitative proteomics principle on high-resolution mass spectrometers: the instrument isolates a scheduled precursor, fragments it, and records a full MS2 spectrum so chosen fragment ions can be extracted after acquisition for relative or absolute peptide quantification (Peterson et al. 2012). Unlike MRM or SRM on a triple quadrupole, which predefines Q3 transitions during acquisition, PRM keeps post-run ion selection flexible while still requiring a closed peptide list, retention-time scheduling, and matched sample prep. Intensities remain relative unless stable isotope-labeled standards and calibration are added (Lange et al. 2008). Prefer PRM when targets are known and high-resolution MS2 specificity helps; keep discovery DIA or LFQ open when the inventory is still changing.
Researchers who already have a closed peptide panel or a specific validation objective can also review the MtoZ Biolabs Parallel Reaction Monitoring (PRM) Service and Targeted Proteomics Service for project-specific feasibility and assay planning.
Principle: What PRM Records and Why It Matters
PRM answers a closed-list question. Only precursors placed on the inclusion list are isolated. Each isolation event yields a full high-resolution product-ion spectrum. Quantification uses extracted fragment chromatograms; qualifier ions support identity. The principle therefore separates three layers that must stay distinct:
Layer |
What it means in PRM |
Common misread |
|---|---|---|
Targeting rule |
Only scheduled peptides are measured |
PRM surveys an open proteome |
Spectral record |
Full MS2 stored for each scheduled precursor |
Every fragment must be locked before the run like MRM |
Quantitative claim |
Relative same-peptide areas by default |
Absolute concentration without SIS and calibration |
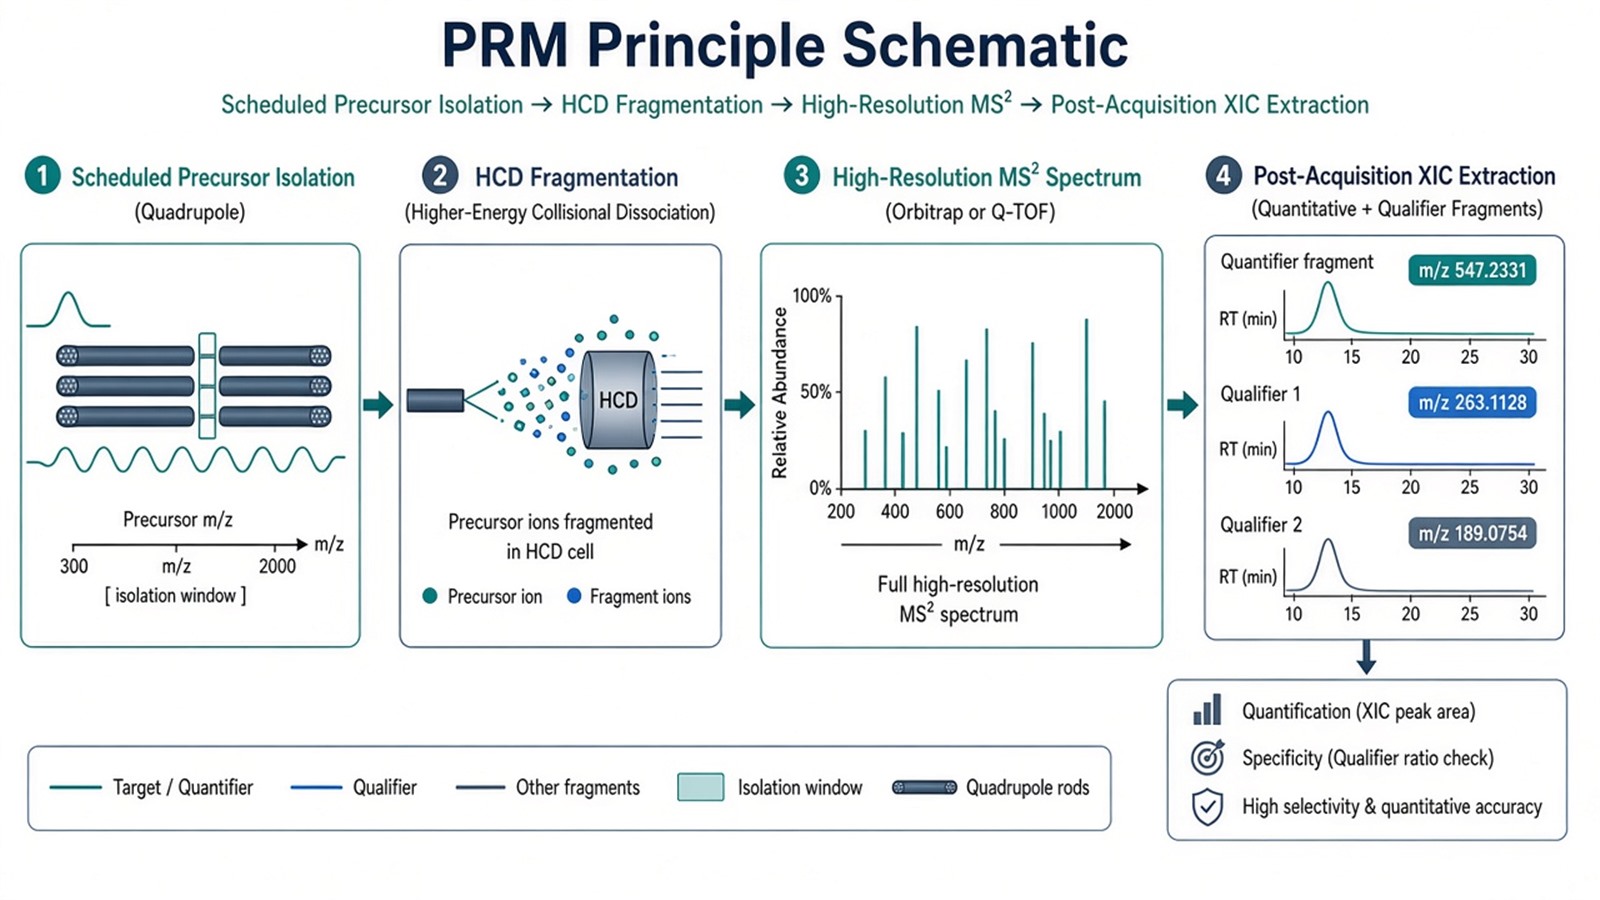
Figure 1. PRM principle: isolate a scheduled precursor, record a full high-resolution MS2 spectrum, and extract quantitative fragment chromatograms after the run.
If a precursor was never scheduled, PRM will not quantify it, regardless of abundance in the digest.
The isolation window is still a window. Co-isolated ions can fragment with the target. Accurate-mass detection reduces many interferences; it does not invent precursor uniqueness when overlapping products remain (Peterson et al. 2012).
How the PRM Principle Differs From MRM and DIA
Method |
Acquisition principle |
Typical fit |
|---|---|---|
PRM |
Scheduled precursor; full HR MS2; post-run fragment choice |
Closed list needing accurate-mass fragment review |
MRM / SRM |
Scheduled precursor-to-product transitions on a triple quadrupole |
Mature, high-multiplex transition panels |
DIA / SWATH |
Wide windows fragment many precursors; library or library-free extraction |
Open or semi-open relative discovery |
PRM is not “DIA with better specificity by name.” DIA remains a discovery-oriented windowed method. PRM remains a closed inclusion-list method. Adding peptides to a PRM panel still requires re-scheduling cycle time; it does not recreate proteome-wide coverage.
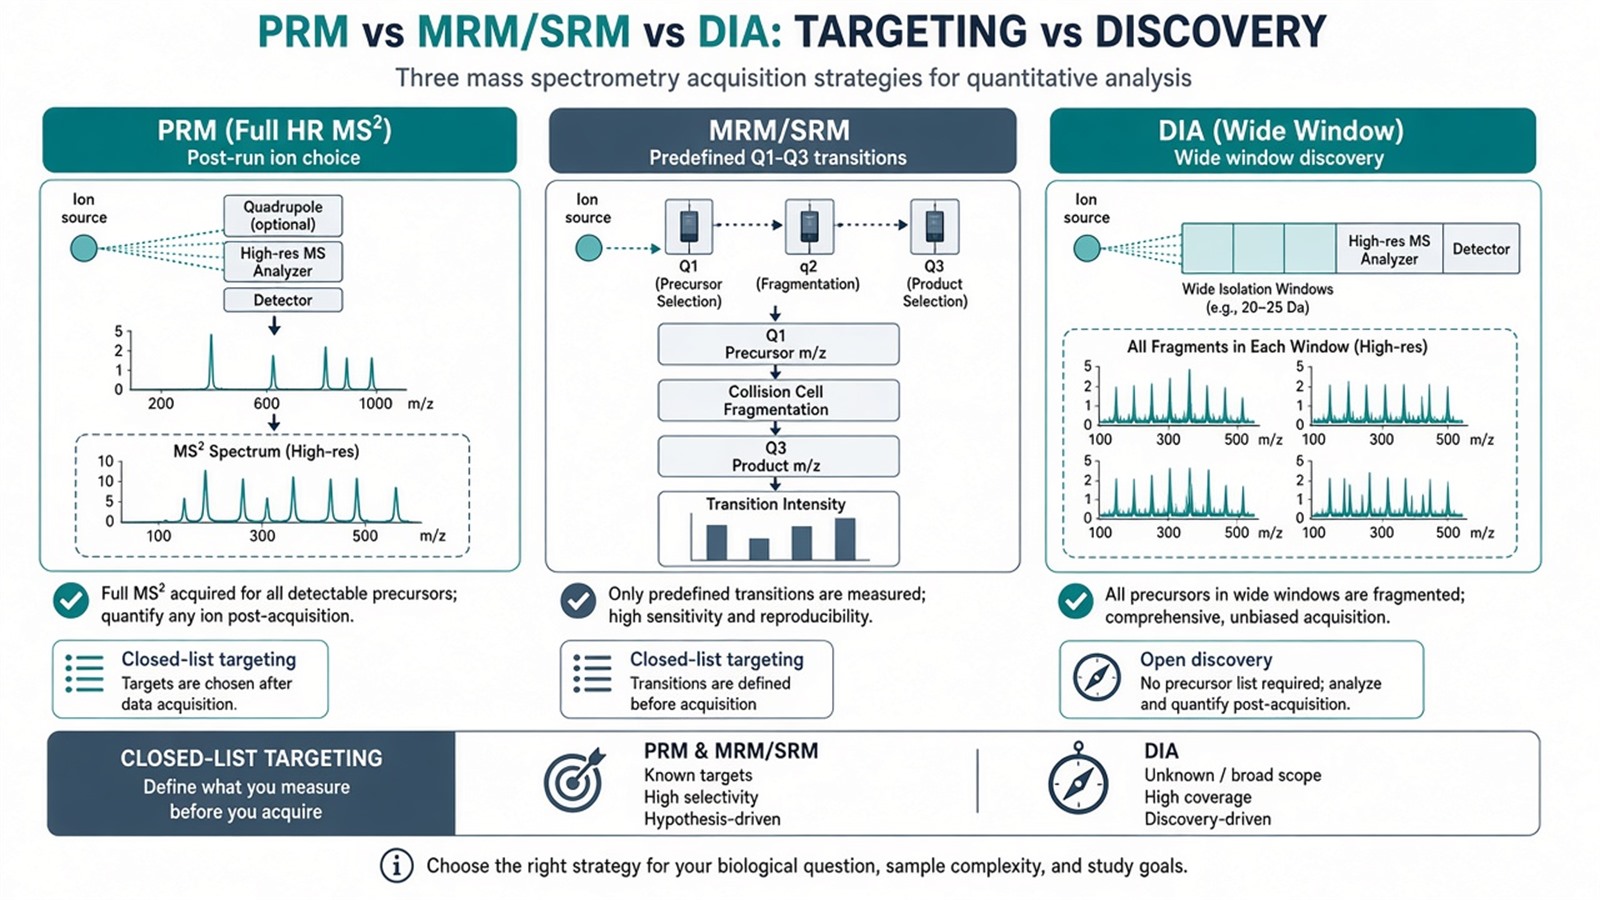
Figure 2. PRM shares closed-list targeting with MRM but records full high-resolution MS2; it does not replace DIA discovery.
For operational workflow beyond the principle, see Parallel Reaction Monitoring (PRM): Principles, Workflow, and Applications. For PRM versus MRM selection, see PRM vs MRM: Comparative Analysis in Quantitative Proteomics and PRM and MRM in Targeted Proteomics: Advantages and Limitations. For MRM or SRM principle context, see Principle of Multiple Reaction Monitoring and SRM Mass Spectrometry.
Judgment Criteria: When the PRM Principle Fits
Study need |
PRM often fits |
Reassess before building the inclusion list |
|---|---|---|
Claim |
Relative confirmation of named peptides |
Open discovery inventory still changing |
Evidence need |
Accurate-mass fragment review in complex digests |
Only a locked triple-quadrupole panel is available |
Panel size |
Short to moderate scheduled list |
Very large multiplex that exceeds cycle-time budget |
Absolute amount |
Optional SIS and matrix-matched calibration planned |
Amount required without standards or reserved aliquots |
Upstream data |
DDA or DIA already nominated proteotypic peptides |
No usable peptide candidates yet |
Prefer PRM when targets are closed and high-resolution MS2 flexibility helps method refinement. Prefer MRM when transitions are already validated and high multiplex on a triple quadrupole is the bottleneck. Prefer DIA or LFQ when the protein inventory must stay open. Prefer AQUA-style SIS designs when concentration units are required on a mature shortlist.
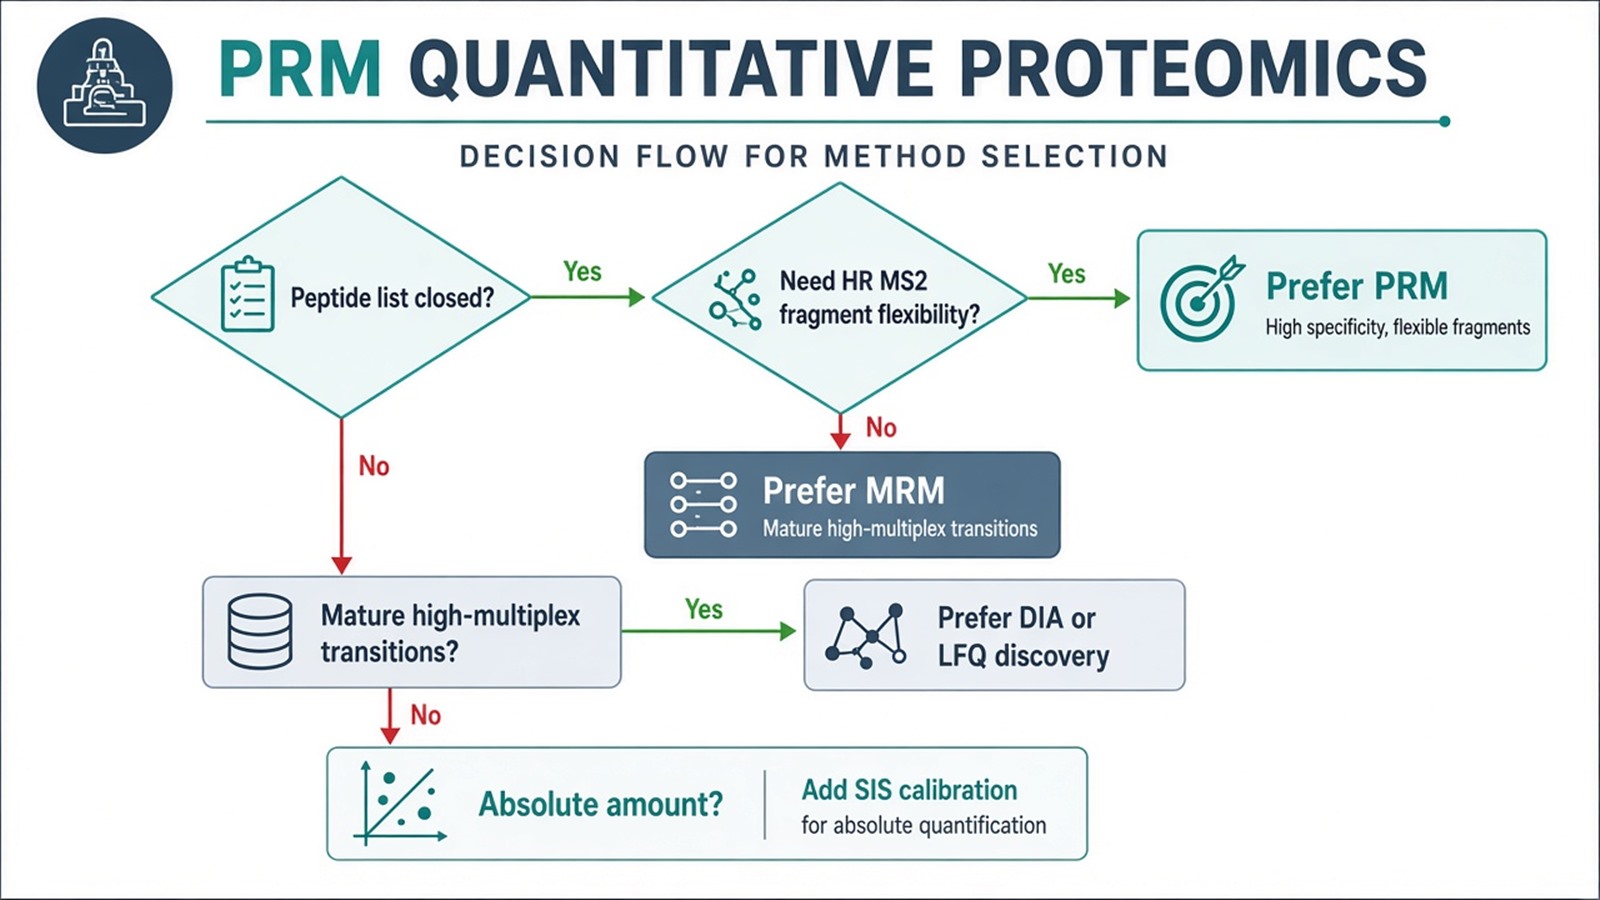
Figure 3. Apply the PRM principle only after the peptide list and claim type are closed; otherwise choose discovery or another targeted route.
For absolute unit design after the principle is clear, see Absolute Quantification in Targeted Proteomics and AQUA Proteomics. For biomarker-validation use of PRM, see How PRM Proteomics Supports Biomarker Validation.
Limits and Application Boundaries
PRM does not measure unscheduled proteins. Full MS2 storage does not remove isolation-window co-fragmentation. Scheduling too many precursors collapses chromatographic sampling and destabilizes peak areas. Relative fragment areas are not molar concentrations without matched standards and calibration. Pathway enrichment from a confirmed shortlist still shows overrepresentation relative to that panel, not whole-proteome pathway proof.
Typical applications aligned with the principle:
- Confirming discovery candidates with proteotypic peptides in complex matrices.
- Pathway or pharmacodynamic panels where fragment identity must be reviewed.
- Selected modified-peptide checks when discriminating ions can be defined.
- Absolute follow-up when SIS peptides and calibration are added to the same closed list.
Diagnostic cutoffs and companion-diagnostic claims sit outside ordinary research PRM reporting unless a separately validated assay framework is in place.
Once the peptide list, relative-versus-absolute claim, and scheduling budget are defined, the project can be scoped as a PRM quantitative study. Researchers can review the MtoZ Biolabs Parallel Reaction Monitoring (PRM) Service for sample evaluation and assay planning.
Frequently Asked Questions
1. What is the core principle of PRM?
Isolate scheduled precursors, record full high-resolution MS2 spectra, and quantify from extracted fragment chromatograms after acquisition.
2. Does PRM automatically give absolute quantification?
No. Absolute amount requires matched stable isotope-labeled standards and calibration. Otherwise the claim stays relative.
3. How does PRM differ from MRM in principle?
MRM predefines Q3 transitions during acquisition. PRM stores a full MS2 spectrum and allows post-run fragment choice.
4. Can PRM replace DIA for discovery?
No. PRM only measures the inclusion list. DIA or LFQ remains the usual open discovery route.
5. Why can overpacking the PRM schedule hurt quantification?
Cycle time is finite. Too many precursors reduce points across the chromatographic peak and make areas unstable.
Reference
- A.C. Peterson, J.D. Russell, D.J. Bailey, M.S. Westphall, J.J. Coon (2012). Parallel reaction monitoring for high resolution and high mass accuracy quantitative, targeted proteomics. Mol. Cell. Proteomics, 11, 1475-1488. https://doi.org/10.1074/mcp.O112.020131
- V. Lange, P. Picotti, B. Domon, R. Aebersold (2008). Selected reaction monitoring for quantitative proteomics: a tutorial. Mol. Syst. Biol., 4, 222. https://doi.org/10.1038/msb.2008.61