



SWATH-MS is a data-independent acquisition (DIA) method that steps consecutive precursor m/z windows across an LC run and fragments all ions in each window every cycle, instead of stochastically picking top precursors as in DDA (Gillet et al. 2012). Window spectra are chimeric, so quantification extracts peptide peak groups from fragment chromatograms with a spectral library or defended library-free search under FDR control. SWATH is one DIA implementation, not a separate class beside DIA. The default output is relative same-protein change across matched injections. Advantages such as lower stochastic missingness and retrospective trace review hold only when window plans, libraries, and prep stay matched (Demichev et al. 2020).
Researchers who already have defined samples, comparison groups, or a specific project objective can also review the MtoZ Biolabs Deep DIA Proteomics Service and Label-Free DIA Quantitative Proteomics for project-specific feasibility and analysis planning.
SWATH-MS Principle: Windows, Libraries, and Peak Groups
DDA ranks eluting precursors and triggers MS/MS on a subset. SWATH schedules fixed or staggered windows and records MS/MS for every ion in each window. Identification and quantification therefore depend on targeted extraction of expected fragments with retention-time alignment and interference filters, not on one clean precursor spectrum per peptide (Gillet et al. 2012).
Principle element |
What SWATH records |
What analysis must add |
|---|---|---|
Isolation |
Sequential windows across m/z |
Documented window width and cycle time |
Spectra |
Composite MS2 per window |
Peak-group extraction and FDR control |
Library role |
Expected fragments and RT anchors |
Matrix-matched or library-free route |
Quant feature |
Fragment chromatogram areas |
Normalization across independent runs |
Claim |
Relative discovery by default |
SIS or targeted assays for amount units |
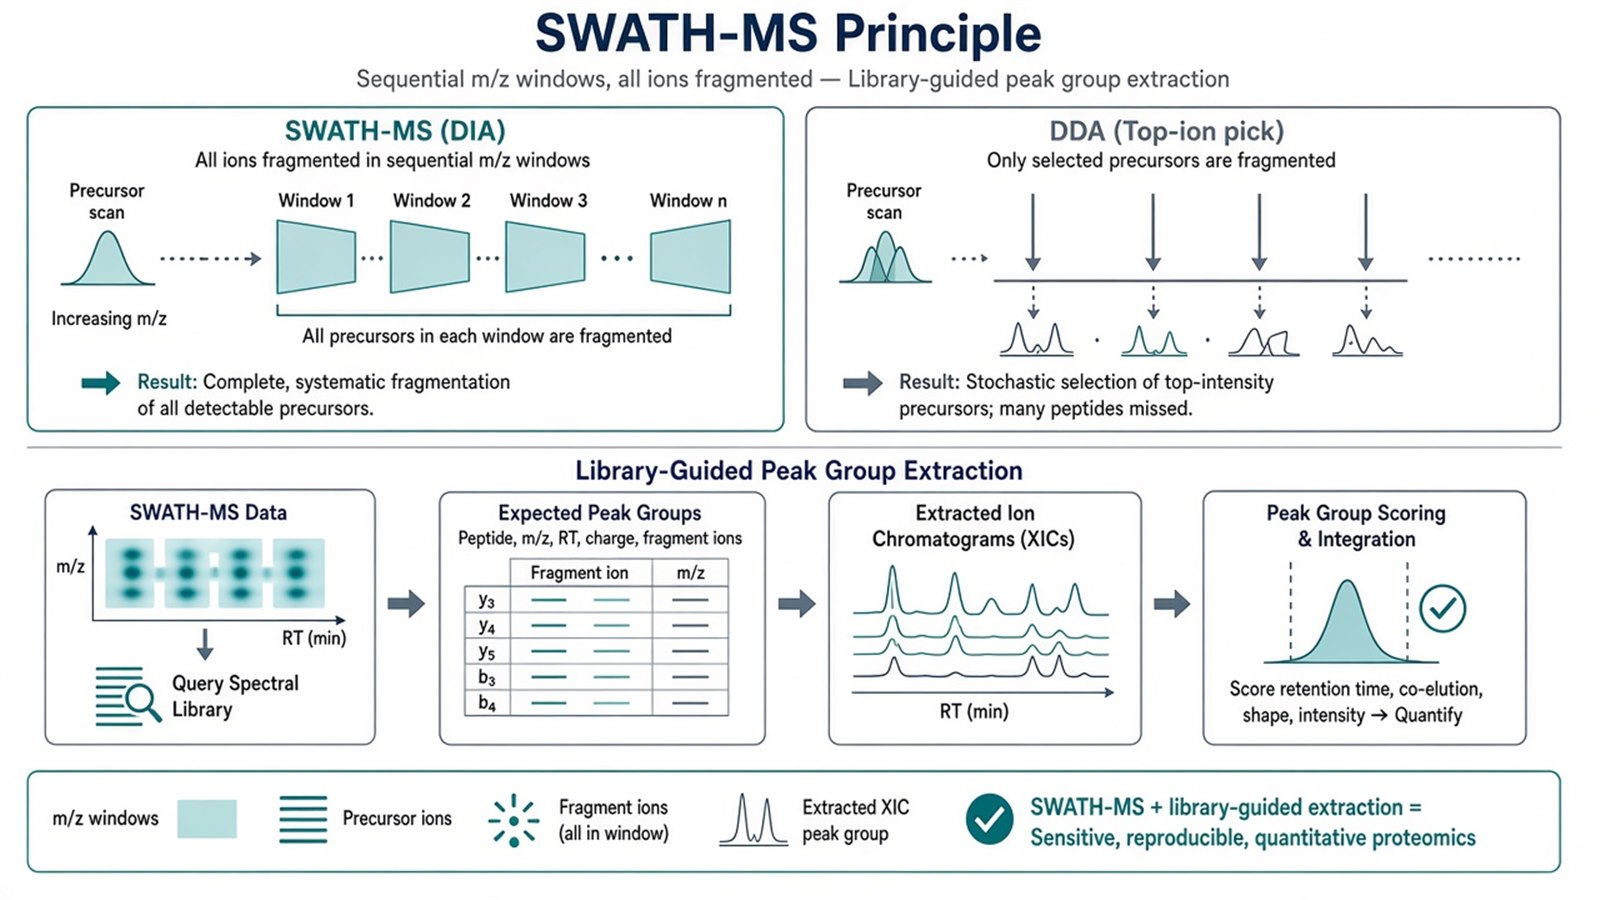
Figure 1. SWATH-MS principle couples windowed acquisition with post-acquisition extraction; it does not assign quantities from raw windows alone.
Calling SWATH comprehensive detection of every peptide in one cycle overstates acquisition; quantified peptides still pass extraction and FDR gates.
For DIA family context, see Principles and Applications of Proteomics Data-Independent Acquisition (DIA) Technology and What Is DIA and SWATH? Understanding Data-Independent Acquisition in Proteomics. For analysis pitfalls, see SWATH-MS Data Analysis Workflow and Common Pitfalls.
Advantages of SWATH-MS and Matching Limits
SWATH advantages and limits arise together. Stating advantages without these limits overstates the technology name alone.
Advantage when QC holds |
Matching limit to keep visible |
|---|---|
More systematic window coverage than stochastic DDA for many peptides |
Co-fragmentation inside each window |
Independent unlabeled enrollment across runs |
Chromatographic and prep batch risk per injection |
Stored windows allow retrospective peptide trace review under FDR |
Wrong library or weak scoring creates false peak groups |
Flexible cohort size without a fixed isobaric channel count |
Instrument time scales with sample number |
Relative discovery matrices with often lower missingness |
Completeness is not accuracy; not absolute concentration by default |
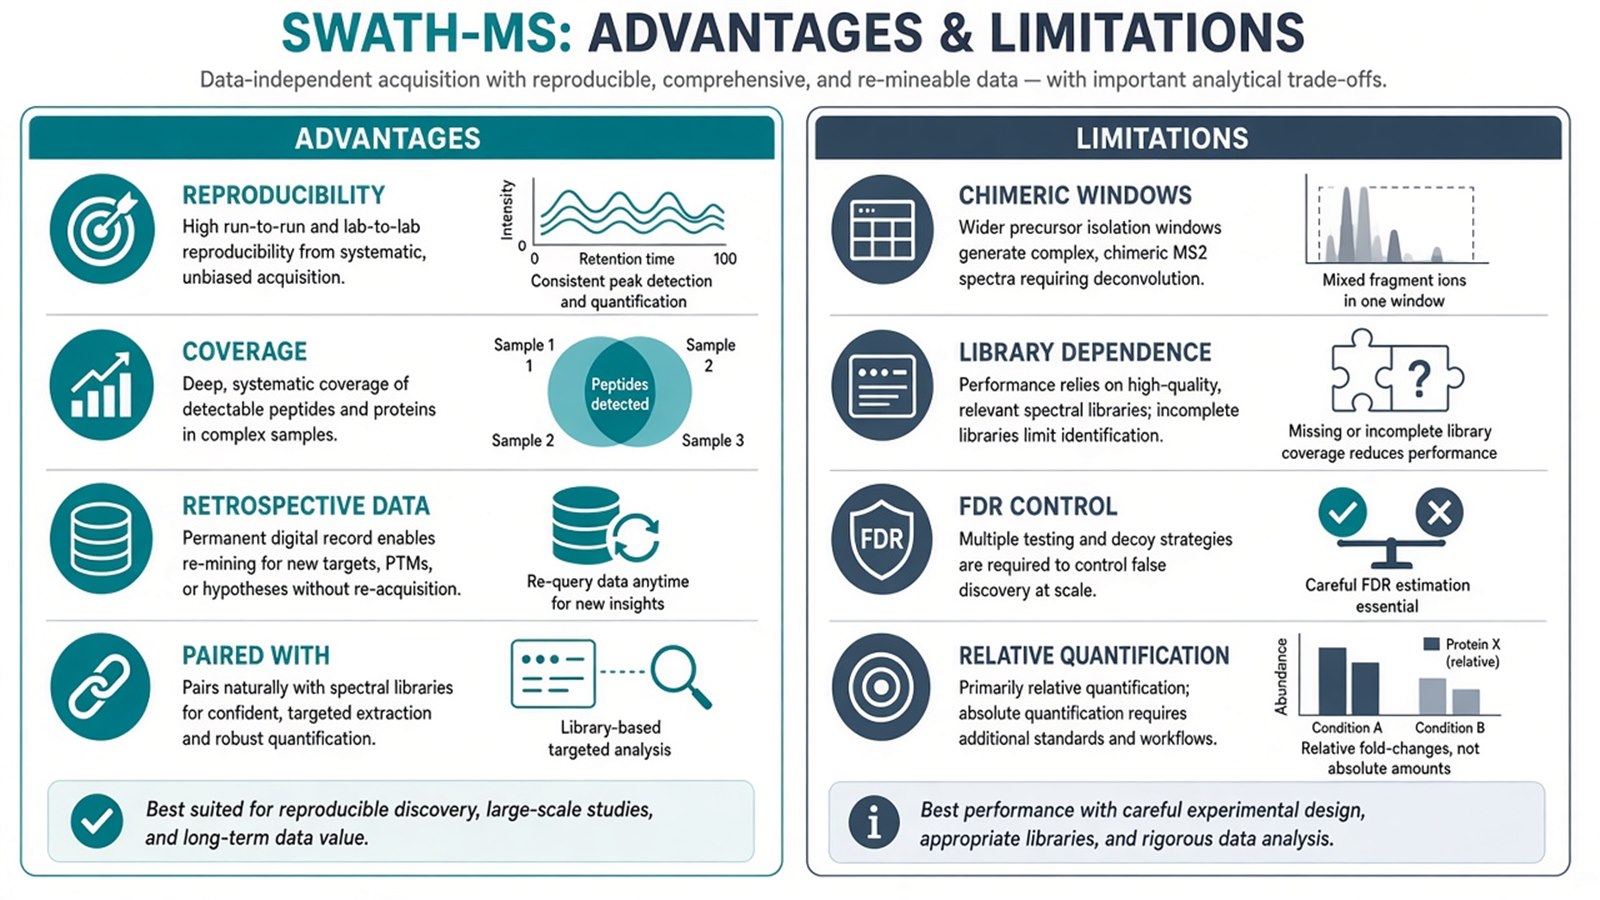
Figure 2. SWATH-MS advantages depend on window design, library or library-free QC, and a declared relative claim.
For full advantage and disadvantage lists, see Advantages and Disadvantages of SWATH-MS and Advantages and Disadvantages of DIA in Protein Quantification. For DDA contrast, see SWATH-MS vs DDA-MS: Which Technique Is Suitable for Your Research?.
Judgment Criteria: When SWATH-MS Technology Fits
Study need |
SWATH-style DIA often fits |
Reassess |
|---|---|---|
Claim |
Relative unlabeled discovery across runs |
Absolute concentration or diagnostic cutoff |
Coverage |
Lower DDA missingness is a priority |
Single-precursor MS2 for novel ID dominates |
Library or extractor |
Matrix-matched library or defended library-free path |
No FDR-controlled extraction capacity |
Enrollment |
Independent injections; samples may arrive later |
All contrasts must share one TMT plex |
Follow-up |
Discovery list may move to PRM or AQUA |
Closed shortlist already needs amount units now |
Prefer SWATH-style DIA when windowed files, extraction QC, and flexible enrollment fit the relative claim. Prefer DDA-LFQ when identification-centric spectra come first. Prefer TMT when same-run multiplex ratios fit a frozen kit. Prefer PRM or AQUA when targets are closed or amount units are required.
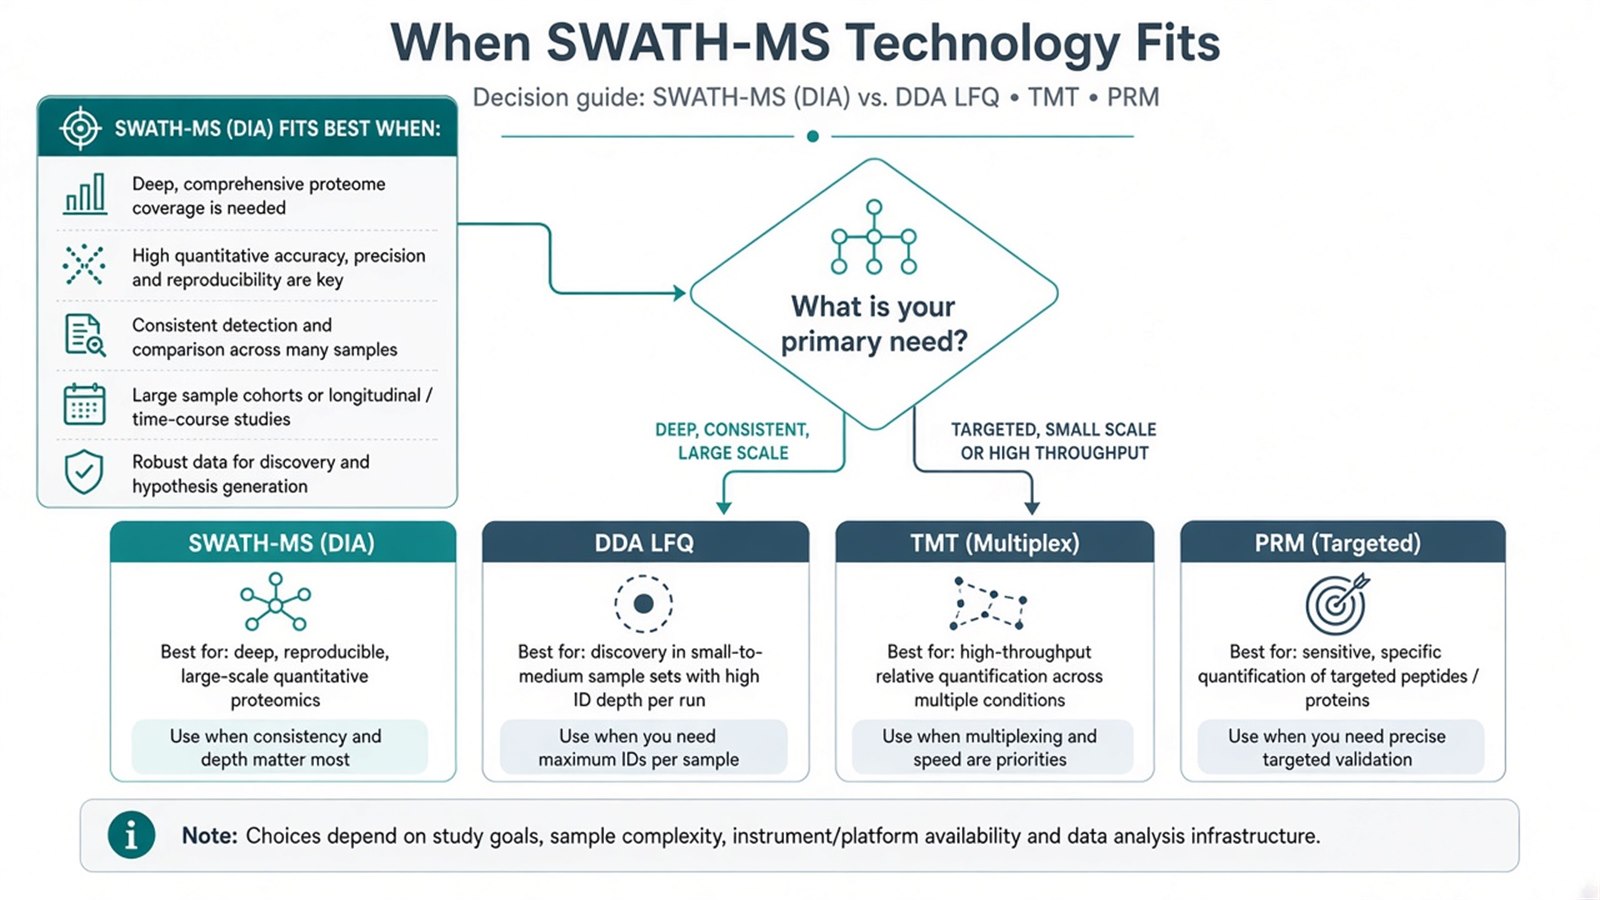
Figure 3. Choose SWATH-MS when DIA window principles and unlabeled relative discovery align; change route when labeling or closed targeted panels dominate.
For strategy context, see DDA vs. DIA vs. TMT in LC-MS/MS Proteomics: How to Choose the Right Strategy and Label-Free Quantification Methods: Mass Spectrometry-Based Protein Quantification Strategies. For library-free options, see Direct DIA: High-Efficiency Proteomics Without a Spectral Library.
Limits and Application Boundaries
SWATH-MS does not remove batch effects without matched prep and run-order control. High-resolution hardware does not erase interference when windows are too wide or libraries mismatch the matrix. Retrospective re-extraction can review stored data but does not guarantee correct peak groups without versioned rules. Pathway enrichment from differential lists shows overrepresentation relative to the quantified background; it does not prove pathway activity. Personalized medicine or diagnostic deployment sits outside ordinary research SWATH reporting unless a separately validated assay framework is in place.
Typical applications when principle, advantages, and limits align:
- Cohort relative discovery with flexible unlabeled enrollment.
- Biofluid or tissue screens where DDA missing values would dominate.
- Mechanism or drug-response studies nominating candidates for PRM confirmation.
- Stored window data revisited for nominated peptides under FDR.
- Large multi-batch designs when window plans and libraries stay version-locked.
Once window scheme, extraction route, and the relative claim are defined, the project can be scoped as a SWATH or DIA study. Researchers can review the MtoZ Biolabs Deep DIA Proteomics Service for sample evaluation and workflow planning.
Frequently Asked Questions
1. Is SWATH-MS the same as all DIA methods?
No. SWATH-MS is a sequential-window DIA implementation, often library-guided. Other DIA window schedules and library-free pipelines share the same data type.
2. What is the main quantitative advantage of SWATH over DDA?
More systematic fragmentation across scheduled windows can reduce stochastic missingness for many peptides when extraction QC holds.
3. Does SWATH-MS eliminate the need for a spectral library?
Classical SWATH workflows often use one. Library-free DIA can analyze similar files but still requires FDR and interference checks.
4. Can SWATH advantages support absolute protein concentration claims?
Not by default. Absolute amount needs matched standards and calibration on a closed panel, usually via targeted assays.
5. When should a project leave SWATH for TMT or PRM?
Choose TMT when a frozen plex fits better. Choose PRM or AQUA when a closed shortlist needs tighter confirmation or amount units.
Reference
- L.C. Gillet, P. Navarro, S. Tate, et al. (2012). Targeted data extraction of the MS/MS spectra generated by data-independent acquisition: a new concept for consistent and accurate proteome analysis. Mol. Cell. Proteomics, 11, O111.016717. https://doi.org/10.1074/mcp.M111.016717
- V. Demichev, C.B. Messner, S.I. Vernardis, K.S. Lilley, M. Ralser (2020). DIA-NN: neural networks and interference correction enable deep proteome coverage in high throughput. Nat. Methods, 17, 41-44. https://doi.org/10.1038/s41592-019-0638-x
