



Label-free quantitative proteomics with sequential window acquisition of all theoretical fragment ion spectra (SWATH-MS) combines unlabeled digests on separate liquid chromatography-tandem mass spectrometry (LC-MS/MS) injections with a windowed data-independent acquisition (DIA) schedule and false discovery rate (FDR) controlled extraction (Gillet et al. 2012). Label-free names the chemistry. SWATH names one common tiled window plan inside DIA. The guide below integrates label-free prep, SWATH acquisition scope, analysis handoff, limits, and project fit for service planning. It is not a software catalog, not a disease application tour, and not a substitute for a signed methods appendix.
Relative contrast across predefined groups remains the default discovery claim unless parallel reaction monitoring (PRM) or stable isotope-labeled internal standard (SIS) designs are declared for follow-up (Gerber et al. 2003).
In MtoZ Biolabs offerings, label-free SWATH-style DIA is commonly evaluated when open contrasts, matched pretreatment, and separate injections fit the cohort. Named peptides may move to targeted remeasurement after discovery when scope includes confirmation.
General DIA service scoping compares with DIA-Based Protein Quantitative Service. SWATH mechanism language compares with Principle of SWATH-Based Protein Quantification.
How Label-Free SWATH Fits a Project Record
Three scopes belong in the brief and stay separated.
Chemistry scope is label-free: no isobaric or metabolic tags on discovery injections. Each sample follows matched prep to an injection-ready digest.
Acquisition scope is SWATH-style DIA: locked m/z windows, cycle time, and gradient family across the cohort block. Mid-study window changes split comparability.
Analysis scope covers library or library-free extraction, unified FDR, normalization, missing-value policy, and contrast testing on frozen groups only (Demichev et al. 2020; Rosenberger et al. 2017).
Label-free SWATH service value is a documented relative matrix for a declared contrast, not unlimited retrospective reanalysis without a new analysis record.
Integrated SWATH planning appears in SWATH Proteomics. Label-free plus 4D or DIA integration appears in 4D-DIA Label-Free Quantitative Proteomics: the Key Technology in Proteomics.
Information to Prepare Before Label-Free SWATH Work Starts
Feasibility review is most useful when fields are explicit.
Sample matrix and storage history for cells, tissues, biofluids, or other inputs.
Contrast map with arms, replicates, and whether enrollment may continue after the first acquisition week.
Pretreatment constraints such as depletion or fractionation that must apply equally to every arm.
Library strategy when library-based extraction is planned: provenance matched to matrix and instrument generation, or a declared library-free route.
Intended claim unit as relative discovery, candidate list for PRM, or amount on named peptides through targeted design.
Matrix prep expectations appear in SWATH-MS Label-Free Protein Quantification Technology Analysis and SWATH MS Analysis. Readiness checks appear in Exploring Quantitative Analysis in Proteomics.
End-to-End Label-Free SWATH Service Workflow
A typical label-free SWATH spine aligns with the following stages.
- Feasibility review of matrix, contrast, and claim unit.
- Matched protein extraction, digestion, and peptide cleanup to LC-ready loads (Wisniewski et al. 2009).
- Pilot LC-MS when the matrix or window plan is new.
- SWATH-style acquisition under a locked plan with interleaved run order and QC inserts as agreed.
- Peptide extraction under declared FDR and interference rules.
- Normalization, missing-value handling, protein rollup, and testing of the predefined contrast only.
- Deliverables within agreed scope, commonly quantitative tables and methods metadata.
- Optional PRM or SIS on a short list when confirmation or amount is in scope.
Steps 2 through 4 define cohort comparability. Step 5 onward follows DIA Data Analysis Methods in Proteomics. Library workflow context appears in Quantitative Analysis of SWATH Proteomics Using OpenSWATH: A Complete Workflow Overview. Operational DIA stages appear in Workflow of DIA in Protein Quantification.
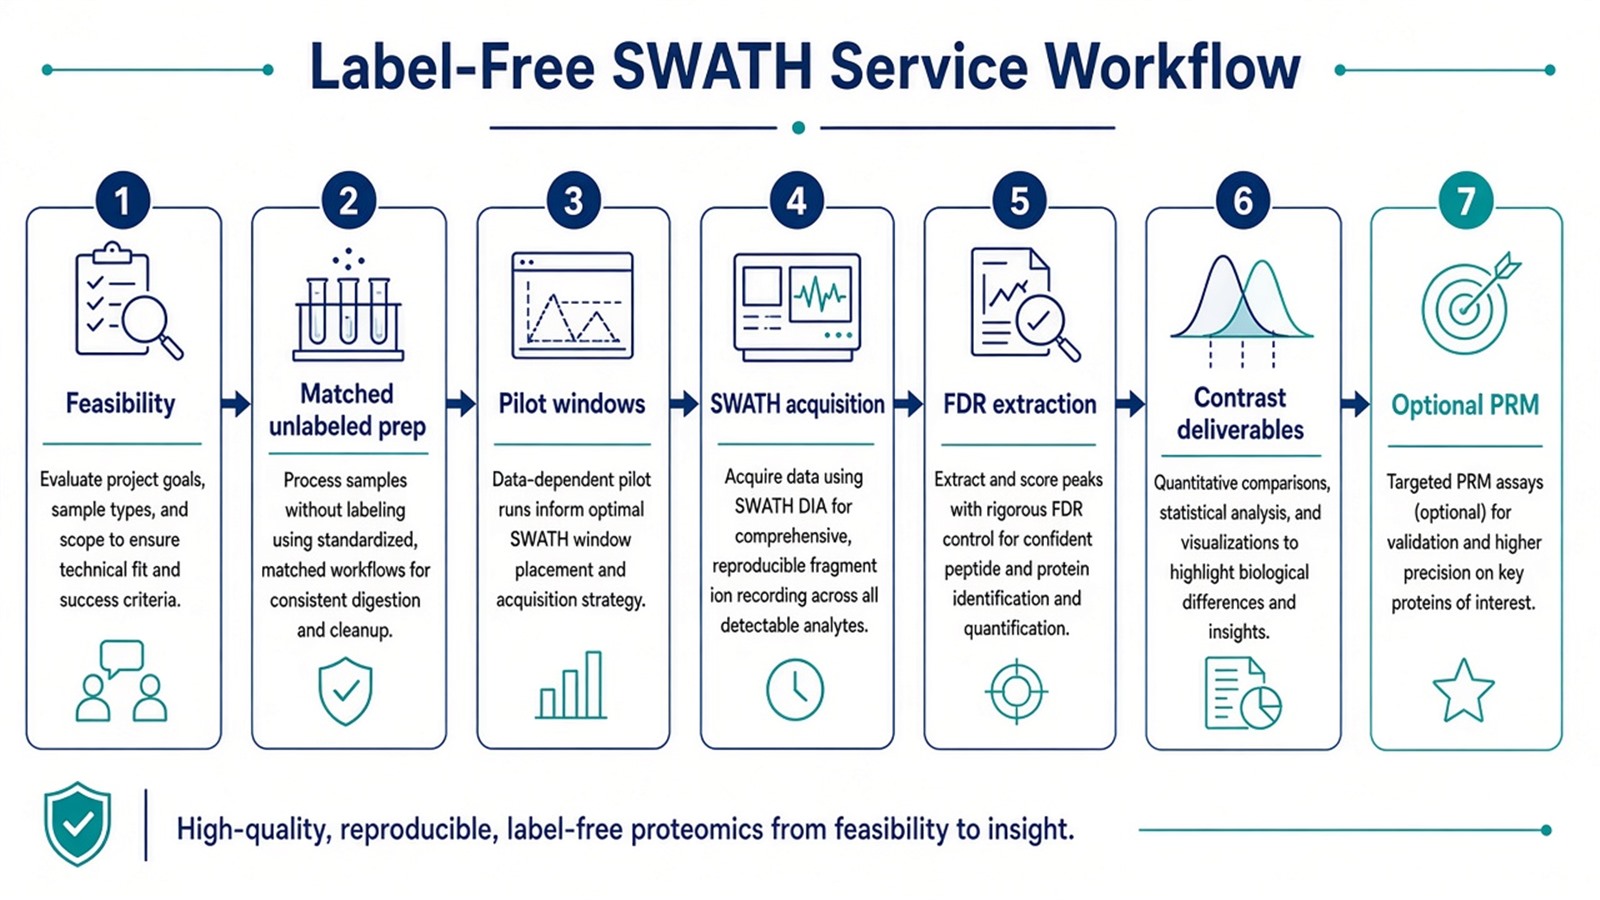
Figure 1. Label-free SWATH projects depend on matched unlabeled prep and a frozen contrast before windowed acquisition and extraction.
Application framing for cohort types appears in Application of SWATH-Based Protein Quantification.
Parameters and Limits in Label-Free SWATH Projects
Scope |
Parameter that must be declared |
Limit if mis-set |
|---|---|---|
Chemistry |
Unlabeled separate injections |
Tagged files merged into LFQ contrast |
Prep |
Matched across arms |
Load bias mimics biology |
Windows |
Width, count, cycle time locked |
Mid-cohort retuning |
Library |
Source matrix and generation |
Wrong expected transitions |
Extraction |
Library-based versus library-free |
Mixed pipelines in one contrast |
FDR |
Unified model across files |
Drifting cutoffs |
Run order |
Interleaving across groups |
Batch tracks treatment |
Claim |
Relative discovery default |
Open intensities as amount |
Limits follow the same table. Label-free SWATH does not certify diagnostic panels or drug labels. Stored SWATH files support reanalysis only under a declared extractor and contrast. Low-abundance proteins may remain missing when abundance and prep limit sampling (Lazar et al. 2016). Forced SWATH when DDA label-free pilots already meet contrast needs may add cost without gain.
High-file cohort context appears in High-throughput SWATH-MS Quantitative Proteomics for Biomarker Discovery.
How to Choose Label-Free SWATH for the Next Project
Use enrollment, matrix pilots, and follow-up unit together.
Project need |
Label-free SWATH plan that often fits |
Recheck before committing |
|---|---|---|
Open cohort, staggered samples |
Unlabeled SWATH separate injections |
Same-day TMT multiplex assumption |
Many files, window pilot stable |
Locked SWATH after pilot |
Generic window width without pilot |
Closed set, one labeling day |
TMT or iTRAQ may fit better |
SWATH when mixing is required |
Simple matrix, clean DDA LFQ pilot |
DDA LFQ without SWATH |
SWATH without completeness gain |
Candidates need remeasurement |
SWATH discovery then PRM |
Discovery p-values as final assay |
Amount on named peptides |
SIS targeted assay |
SWATH intensities as concentration |
Mobility interference in pilot |
4D-DIA after pilot lock |
SWATH when mobility gain is needed |
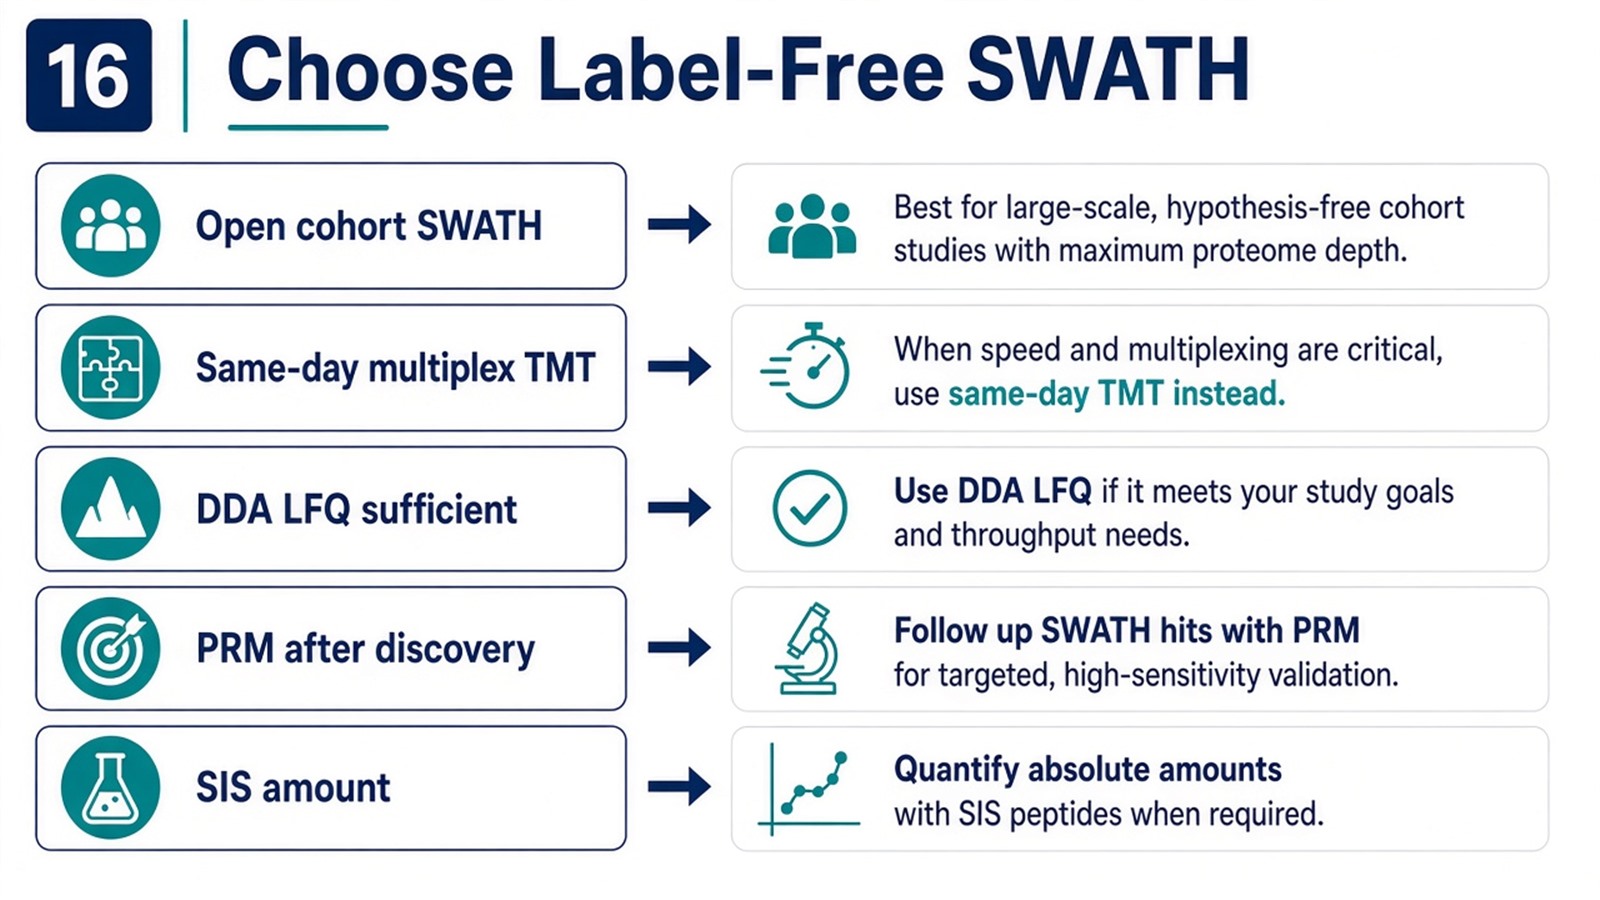
Figure 2. Choose label-free SWATH when unlabeled separate injections and a SWATH window pilot fit the contrast; TMT, DDA LFQ, or PRM fit other patterns.
Three-axis selection appears in Protein Quantification Method. Staged PRM appears in Application of DIA-PRM Proteomics.
Teams with matrix, contrast design, library intent, and follow-up goal can review the MtoZ Biolabs Label-Free Quantitative Proteomics Service for label-free SWATH alignment.
Frequently Asked Questions
1. Is label-free SWATH the same as label-free DDA LFQ?
No. Both use unlabeled separate injections. SWATH is a windowed DIA acquisition plan with fragment-based extraction. DDA LFQ usually uses MS1 areas after intensity-picked fragmentation.
2. Must every SWATH project build a new spectral library?
No. Library-free extraction routes exist when declared in the analysis record. Library-based routes need provenance matched to the matrix.
3. Can SWATH data be re-mined for any new contrast later?
Reanalysis needs a declared extractor and contrast. Relabeling arms after acquisition is not a valid new contrast.
4. Does SWATH service include pathway or multi-omic reporting by default?
Interpretation layers stay within agreed scope. Pathway overlay is optional after contrast QC.
5. When should projects use general DIA windows instead of SWATH tiling?
When pilots show another window layout fits elution density and cycle time on the platform.
MtoZ Biolabs can discuss label-free SWATH project planning when sample type, contrast design, pilot data, and library strategy are documented.
If further evaluation of window pilot lock, extractor choice, or discovery-to-targeted staging is needed, project details can be submitted for a feasibility review.
Reference
- S.A. Gerber, J. Rush, O. Stemman, M.W. Kirschner, S.P. Gygi (2003). Absolute quantification of proteins and phosphoproteins from cell lysates by tandem MS. Proc. Natl. Acad. Sci. U.S.A., 100, 6940-6945. https://doi.org/10.1073/pnas.0832254100
- L.C. Gillet, P. Navarro, S. Tate, H. Rost, H. Selevsek, L. Reiter, R. Bonner, R. Aebersold (2012). Targeted data extraction of the MS/MS spectra generated by data-independent acquisition: a new concept for consistent and accurate proteome analysis. Mol. Cell. Proteomics, 11, O111.016717. https://doi.org/10.1074/mcp.O111.016717
- V. Demichev, C. Messner, S. Vernardis, K.S. Lilley, M. Ralser (2020). DIA-NN: neural networks and interference correction enable deep proteome coverage in high throughput. Nat. Methods, 17, 41-44. https://doi.org/10.1038/s41592-019-0638-x
- G. Rosenberger, S. Liu, J. Röst, H.L. Ludwig, C. Buil, B. Borchers, R. Aebersold, L. Collins (2017). Statistical control of peptide and protein error rates in large-scale targeted data-independent acquisition analyses. Nat. Commun., 8, 14224. https://doi.org/10.1038/ncomms14224
- C. Lazar, L. Gatto, M. Ferro, C. Bruley, T. Burger (2016). Accounting for the multiple natures of missing values in label-free quantitative proteomics data sets to compare imputation strategies. J. Proteome Res., 15, 1116-1125. https://doi.org/10.1021/acs.jproteome.5b00981
- J.R. Wisniewski, A. Zougman, N. Nagaraj, M. Mann (2009). Universal sample preparation method for proteome analysis. Nat. Methods, 6, 359-362. https://doi.org/10.1038/nmeth.1322
