



Four-dimensional data-independent acquisition (4D-DIA) quantitative proteomics here means liquid chromatography-tandem mass spectrometry (LC-MS/MS) with ion mobility separation before fragmentation, so precursors are resolved by retention time, m/z, and mobility before windowed MS/MS (Meier et al. 2020). Relative quantification in 4D-DIA usually comes from extracted fragment intensities on unlabeled digests under false discovery rate (FDR) control, not from isobaric reporter channels (Gillet et al. 2012). The guide below answers when chemical labeling is required, optional, or a different route entirely. It is not a TMT protocol manual, not a vendor mobility marketing page, and not a repeat of 4D-DIA versus 4D-PRM list-status rules.
Labeling is not required for standard label-free 4D-DIA relative quantification on separate injections.
Isobaric tandem mass tag (TMT) or isobaric tags for relative and absolute quantitation (iTRAQ) chemistry is a separate decision from adding ion mobility to DIA. Teams can run 4D-DIA without tags when the contrast is open enrollment, separate injections, and a relative matrix claim. Tags may still fit when a closed multi-arm set can be labeled the same day and mixed before one LC path.
In MtoZ Biolabs offerings, mobility-enhanced DIA discovery is commonly run label-free on unlabeled digests, while multiplexed isobaric work is evaluated as a different chemistry path when sample sets freezable for one labeling day.
How Labeling Relates to 4D-DIA Quantification
Three quantitative ideas are often conflated.
Label-free chemistry means digests stay unlabeled through injection. 4D-DIA compares extracted fragment traces across files after prep and acquisition are matched.
Isobaric labeling means peptides are tagged after digestion so reporter ions support ratio readout within a multiplexed MS/MS event. That workflow is not the default 4D-DIA label-free path and needs its own prep and analysis record.
Stable isotope-labeled internal standard (SIS) spiking targets named peptides for amount claims on targeted assays, not for ordinary open 4D-DIA discovery prep (Gerber et al. 2003).
Mobility adds separation before fragmentation. It does not replace the need to declare chemistry, contrast, and claim unit together.
Researchers who need open discovery versus closed 4D-PRM confirmation can refer to 4D-DIA vs 4D-PRM: Which 4D Quantitative Approach Suits Your Project Better. For three-axis route selection across labeling and acquisition, see Protein Quantification Method.
Label-Free versus Isobaric Routes on Mobility-Enhanced DIA
Label-free 4D-DIA more often fits when cohorts may still add samples, contrasts need separate injections, or enrollment spans weeks. Extraction compares intensities across runs after normalization and interference review (Demichev et al. 2020).
Isobaric TMT or iTRAQ more often fits when arms are frozen, material can be labeled in one batch, and digests will share one mixed LC path. Mobility-enhanced acquisition may still appear on some platforms, but the quantitative readout follows reporter rules, not open label-free extraction alone.
Mixed designs need explicit documentation. Labeling one arm only, or merging tagged and untagged files in one contrast, breaks comparability even when mobility settings match.
4D-DIA label-free work still requires matched pretreatment and a predefined contrast, not a mobility keyword alone.
General label-free pretreatment before DIA appears in SWATH-MS Label-Free Protein Quantification Technology Analysis. Extraction and FDR control belong with DIA Data Analysis Methods in Proteomics.
Shared Workflow From Sample to 4D-DIA Matrix
A practical label-free 4D-DIA spine usually runs as follows.
- Confirm matrix, contrast, and relative claim unit.
- Prepare matched digests with shared lysis, digestion, and cleanup rules.
- Desalt to injection-ready loads without isobaric tagging when label-free 4D-DIA is chosen.
- Acquire mobility-aware windowed DIA under a locked plan across the cohort.
- Extract peptides by library or library-free search under FDR and interference rules (Rosenberger et al. 2017).
- Normalize, roll up proteins, and test only the predefined contrast.
- Optional targeted PRM or SIS follow-up on a short list when confirmation or amount is required.
Steps 2 and 3 are where labeling would enter on an isobaric route instead of step 3 staying tag-free. Step 4 onward assumes separate unlabeled injections for label-free 4D-DIA.
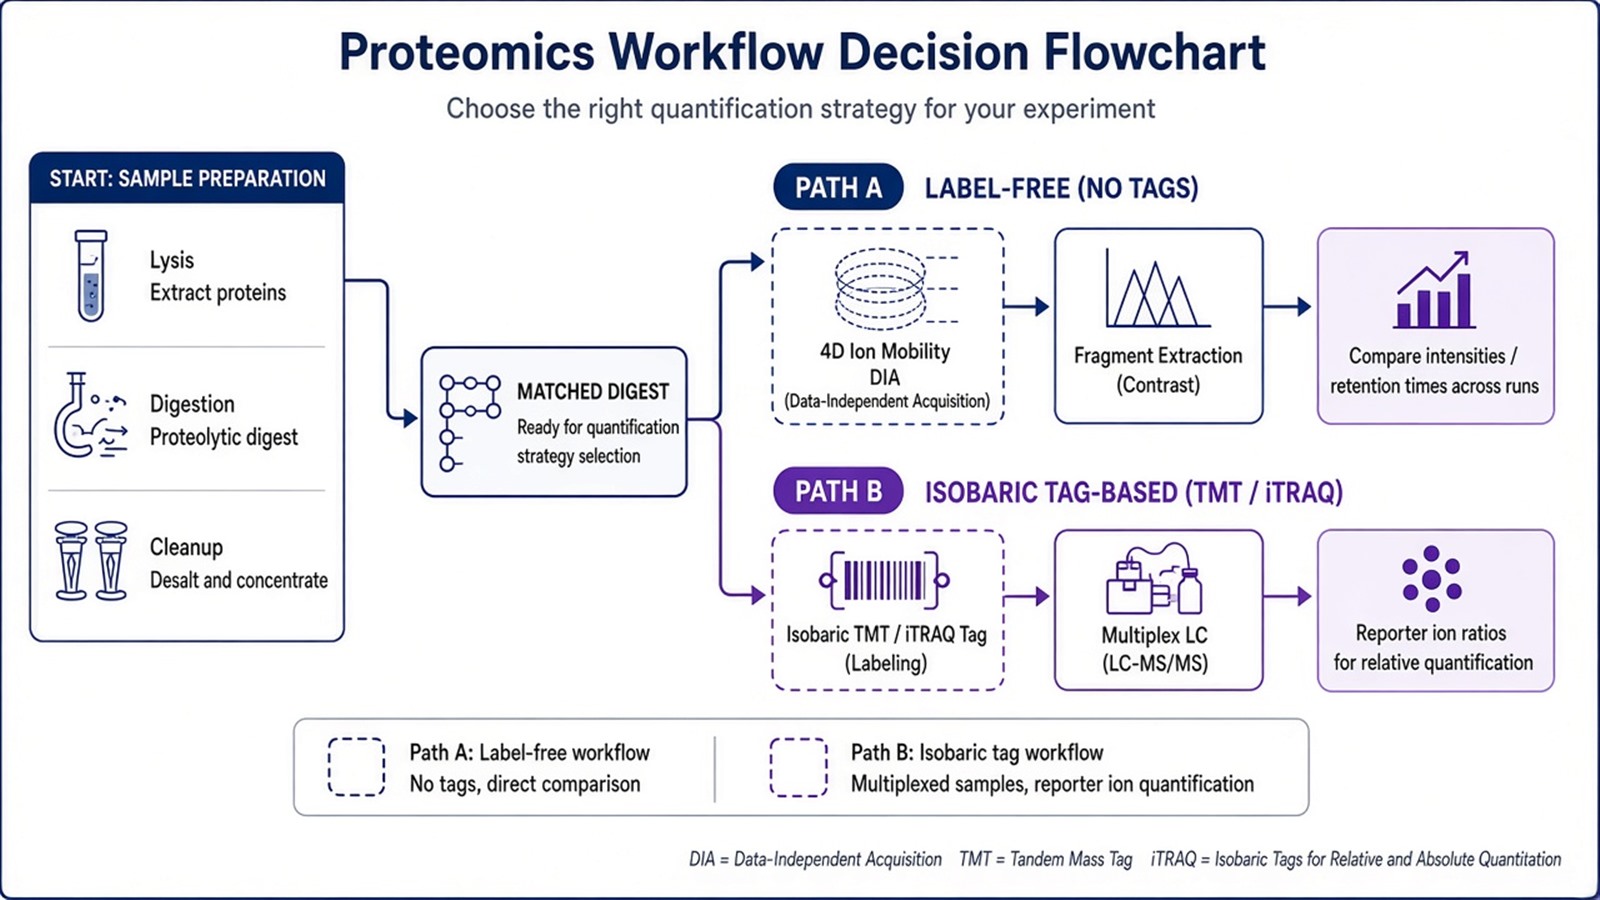
Figure 1. Label-free 4D-DIA skips isobaric tagging before injection; isobaric routes need a closed set and a reporter-based analysis record.
Matched prep spine detail appears in Quantitative Protein Analysis.
Parameters and Limits That Affect the Labeling Decision
Parameter |
Label-free 4D-DIA sensitivity |
Isobaric route sensitivity |
|---|---|---|
Enrollment |
Open cohorts, staggered samples |
Closed set, one labeling day |
Injection path |
Separate digests per sample |
Often multiplexed LC after tagging |
Quant readout |
Extracted fragment intensities |
Reporter ion ratios within multiplex |
Typical claim |
Relative matrix across files |
Relative ratios within TMT/iTRAQ set |
Prep constraint |
Matched cleanup across arms |
Tagging completeness, channel balance |
Follow-up |
PRM on short list after discovery |
May still need targeted confirmation |
Limits follow the same table. Label-free 4D-DIA does not deliver absolute molar concentration without SIS on named peptides. Isobaric tagging does not remove batch effects outside the multiplex design. Mobility improves separation for label-free 4D-DIA. It does not fix unequal prep or undefined contrasts. Switching from label-free to TMT mid cohort invalidates the matrix even when raw files are stored.
How to Choose Labeling for the Next 4D-DIA Project
Use enrollment, claim unit, and list status together.
Project need |
Route that often fits |
Recheck before committing |
|---|---|---|
Open inventory, relative contrast |
Label-free 4D-DIA |
Forced TMT without freezable set |
Samples arrive over weeks |
Label-free separate injections |
Same-day labeling assumption |
Six to sixteen arms, one harvest day |
TMT or iTRAQ multiplex |
Open enrollment on tags |
Confirmation on named peptides |
Label-free 4D-DIA then PRM |
Discovery reporters as final assay |
Amount on selected peptides |
Targeted SIS assay |
Label-free intensities as concentration |
Pilot with unknown contrast |
Label-free pilot, freeze prep SOP |
Full TMT batch before contrast lock |
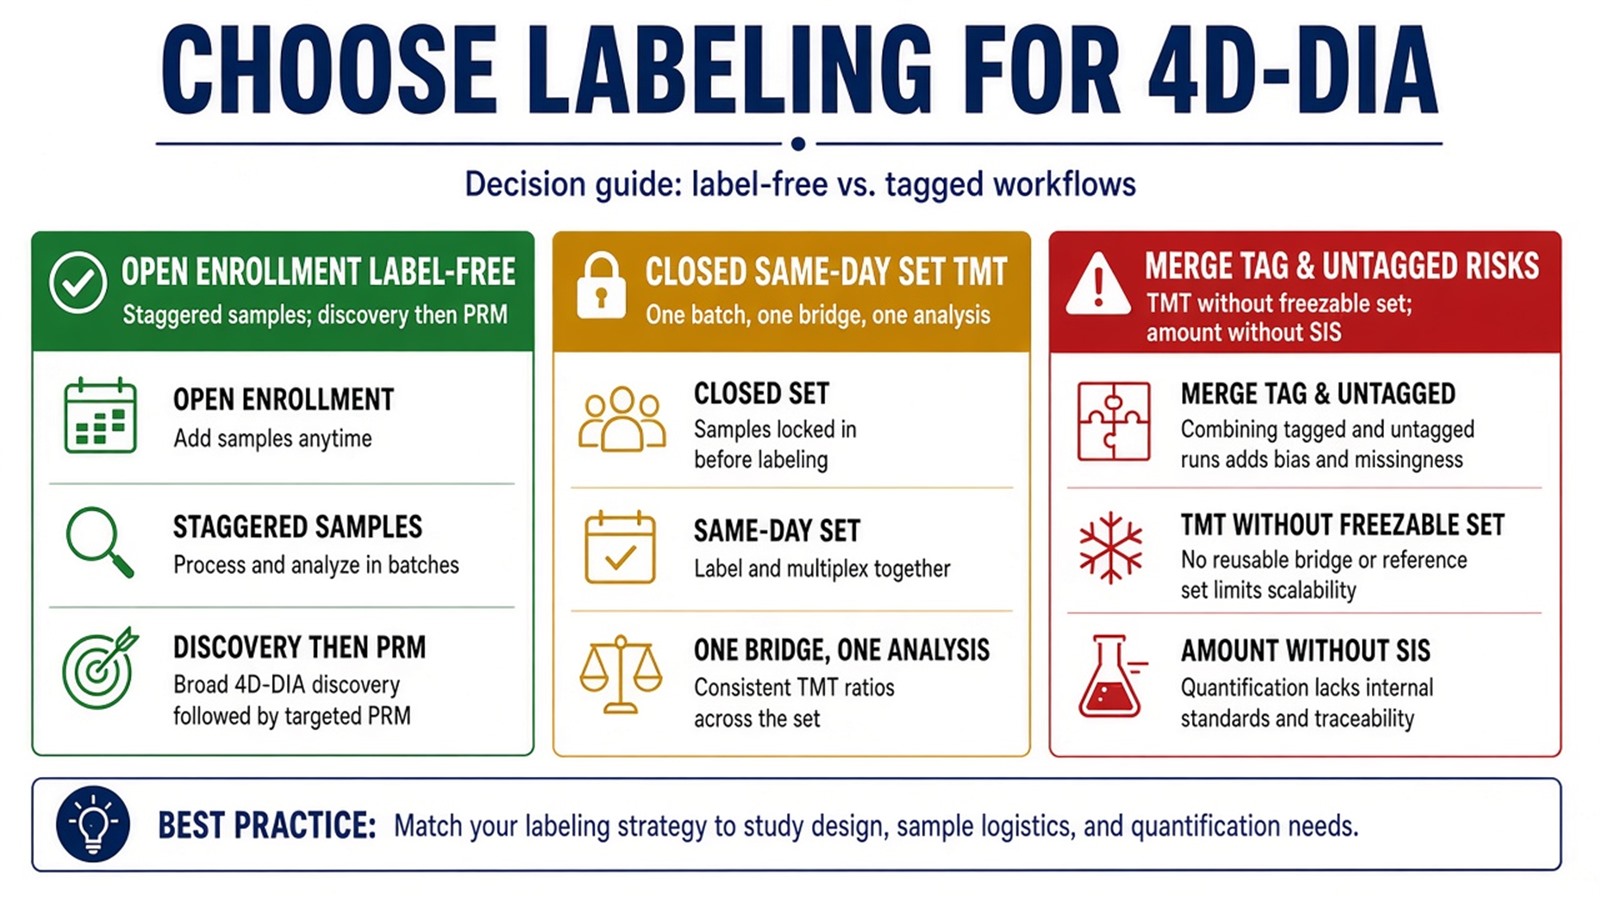
Figure 2. Label-free 4D-DIA fits open contrasts on unlabeled injections; isobaric labeling fits closed same-day sets with reporter readout.
Teams that already have matrix, contrast, enrollment plan, and claim unit can review the MtoZ Biolabs Label-Free Quantitative Proteomics Service for mobility-enhanced DIA planning.
Frequently Asked Questions
1. Is labeling required for 4D-DIA?
No for standard label-free relative 4D-DIA on separate injections. Isobaric labeling is optional when a multiplexed reporter design fits the cohort.
2. Can TMT and label-free 4D-DIA files be merged in one contrast?
No. Chemistry and readout differ. Contrasts should stay within one quantitative route.
3. Does ion mobility remove the need for matched prep?
No. Mobility adds separation before MS/MS. Matched extraction and digestion still gate comparability.
4. Is iTRAQ interchangeable with TMT for this decision?
Both are isobaric routes with similar project-fit rules. Channel count and prep SOP still need declaration.
5. Does label-free 4D-DIA replace PRM validation?
No. Open 4D-DIA supports discovery contrasts. Closed remeasurement is a targeted follow-up design.
MtoZ Biolabs can support 4D-DIA planning when sample type, contrast, enrollment flexibility, and the intended relative or amount claim are available.
If further evaluation of label-free versus isobaric enrollment, mobility window plans, or discovery-to-targeted staging is needed, project details can be submitted for a feasibility review.
Reference
- S.A. Gerber, J. Rush, O. Stemman, M.W. Kirschner, S.P. Gygi (2003). Absolute quantification of proteins and phosphoproteins from cell lysates by tandem MS. Proc. Natl. Acad. Sci. U.S.A., 100, 6940-6945. https://doi.org/10.1073/pnas.0832254100
- L.C. Gillet, P. Navarro, S. Tate, H. Rost, N. Selevsek, L. Reiter, R. Bonner, R. Aebersold (2012). Targeted data extraction of the MS/MS spectra generated by data-independent acquisition: a new concept for consistent and accurate proteome analysis. Mol. Cell. Proteomics, 11, O111.016717. https://doi.org/10.1074/mcp.O111.016717
- F. Meier, A.D. Brunner, O. Frank, M.T. Krauss, M. Koch, H. Teichert, C. Trede, J. Kosinski, T. Park, M. The, M. Mann (2020). diaPASEF: parallel accumulation-serial fragmentation combined with data-independent acquisition. Nat. Methods, 17, 1229-1236. https://doi.org/10.1038/s41592-020-00998-0
- V. Demichev, C. Messner, S. Vernardis, K.S. Lilley, M. Ralser (2020). DIA-NN: neural networks and interference correction enable deep proteome coverage in high throughput. Nat. Methods, 17, 41-44. https://doi.org/10.1038/s41592-019-0638-x
- G. Rosenberger, S. Liu, J. Röst, H.L. Ludwig, C. Buil, B. Borchers, R. Aebersold, L. Collins (2017). Statistical control of peptide and protein error rates in large-scale targeted data-independent acquisition analyses. Nat. Commun., 8, 14224. https://doi.org/10.1038/ncomms14224
