



From TMT to SWATH, quantitative proteomics technology choice is a project-design problem, not a timeline of which method replaced the other. Tandem mass tags (TMT) remain a multiplex isobaric labeling route: digests receive channel-specific tags, are mixed, and report relative ratios from reporter ions on liquid chromatography-tandem mass spectrometry (LC-MS/MS) (Thompson et al. 2003). Sequential window acquisition of all theoretical mass spectra (SWATH) is a data-independent acquisition (DIA) label-free route that fragments scheduled m/z windows and extracts peptide quantities without chemical multiplexing (Gillet et al. 2012). In project language, teams keep TMT when a locked plex and within-mix relative claim fit; they move to SWATH-style DIA when unlabeled injections, expandable cohorts, or library-based reuse fit better. The guide below centers TMT comprehensive scope, workflow, locked parameters, limits, and fit versus SWATH. It is not a 2D-gel history review, not an artificial intelligence multi-omics brochure, and not a claim that SWATH automatically succeeds TMT.
Ratio compression remains a known MS2 TMT limit; MS3 or synchronous precursor selection MS3 (SPS-MS3) can reduce co-isolation mixing on compatible instruments (Ting et al. 2011; McAlister et al. 2014). SWATH still needs matched prep, window design, and library or library-free rules before reproducibility claims hold (Bantscheff et al. 2012).
In MtoZ Biolabs quantitative offerings, TMT-first projects are scoped after plex map, matrix, relative claim, and whether a SWATH or targeted follow-up is planned. Service intake for the labeled route appears in TMT Quantitative Proteomics Analysis Service.
What a TMT-Centered Project Must Cover Before SWATH Is Considered
Four TMT layers belong in the brief even when SWATH is an alternative on the table.
Contrast layer freezes groups, replicates, and the relative claim. “Evolution of platforms” is not a contrast.
Plex-and-channel layer names kit size, channel assignment, bridges for multi-plex batches, and randomization. Channel count is not a biological result.
Prep-and-label layer names matched protein input, digest, labeling completeness, quench, and cleanup. Incomplete labeling breaks reporters before any SWATH comparison matters.
Acquisition-and-hand-off layer names MS2 versus MS3 reporters, fractionation, and whether discovery hits will later move to SWATH depth checks or closed-panel multiple reaction monitoring (MRM) / parallel reaction monitoring (PRM).
TMT answers within-plex relative ratios after a locked channel map. SWATH answers label-free DIA profiles on separate injections. One does not inherit the other’s claim unit.
Plex design and labeling gates appear in Protein Quantification Technology-TMT Labeling Quantitation. This page stays on comprehensive TMT decision framing when SWATH is the main alternative.
When SWATH Becomes the Better Companion or Replacement
SWATH fits when TMT multiplex chemistry is the wrong constraint.
Keep TMT when many samples must share one chromatographic path inside a kit, when reagent cost and channel planning are acceptable, and when the question is relative change inside that mix.
Prefer SWATH-type DIA when samples must stay unlabeled, when the cohort may grow beyond one plex without bridge redesign, or when retrospective window re-extraction matters more than reporter chemistry. Label-free DIA service framing appears in Label-Free DIA Quantitative Proteomics. SWATH-specific intake appears in SWATH.
SWATH does not remove batch prep drift. Library quality, window scheme, and injection order still lock the quantitative record. TMT does not become outdated because DIA exists; the routes remain complementary.
End-to-End Workflow: Lock TMT First, Then Decide on SWATH
A practical TMT-centered spine that can branch to SWATH usually runs as follows.
- Freeze contrast, biological n, and relative claim for the protein quantification question.
- Ask whether samples must share one mixed LC-MS/MS path or remain on separate unlabeled injections.
- If multiplex mixing fits, draft the TMT channel map, bridges, and labeling completeness checks.
- Match protein input and digest; label, quench, and mix only after pilot completeness passes.
- Acquire MS2 or MS3 reporters with isolation settings locked from the pilot; fractionate if depth is required.
- Normalize reporters and test only the predefined contrast; review ratio-compression flags.
- If unlabeled growth, library reuse, or injection-level cohort expansion is required instead, scope SWATH/DIA with matched prep and window or library rules.
- Do not merge TMT reporter matrices and SWATH peak matrices into one undeclared statistical object.
- Hand named peptides to MRM/PRM only when a closed verification claim is declared after either discovery route.
Steps 1 through 6 protect the TMT record. Steps 7 through 9 prevent “evolution” language from rewriting claim units after acquisition.
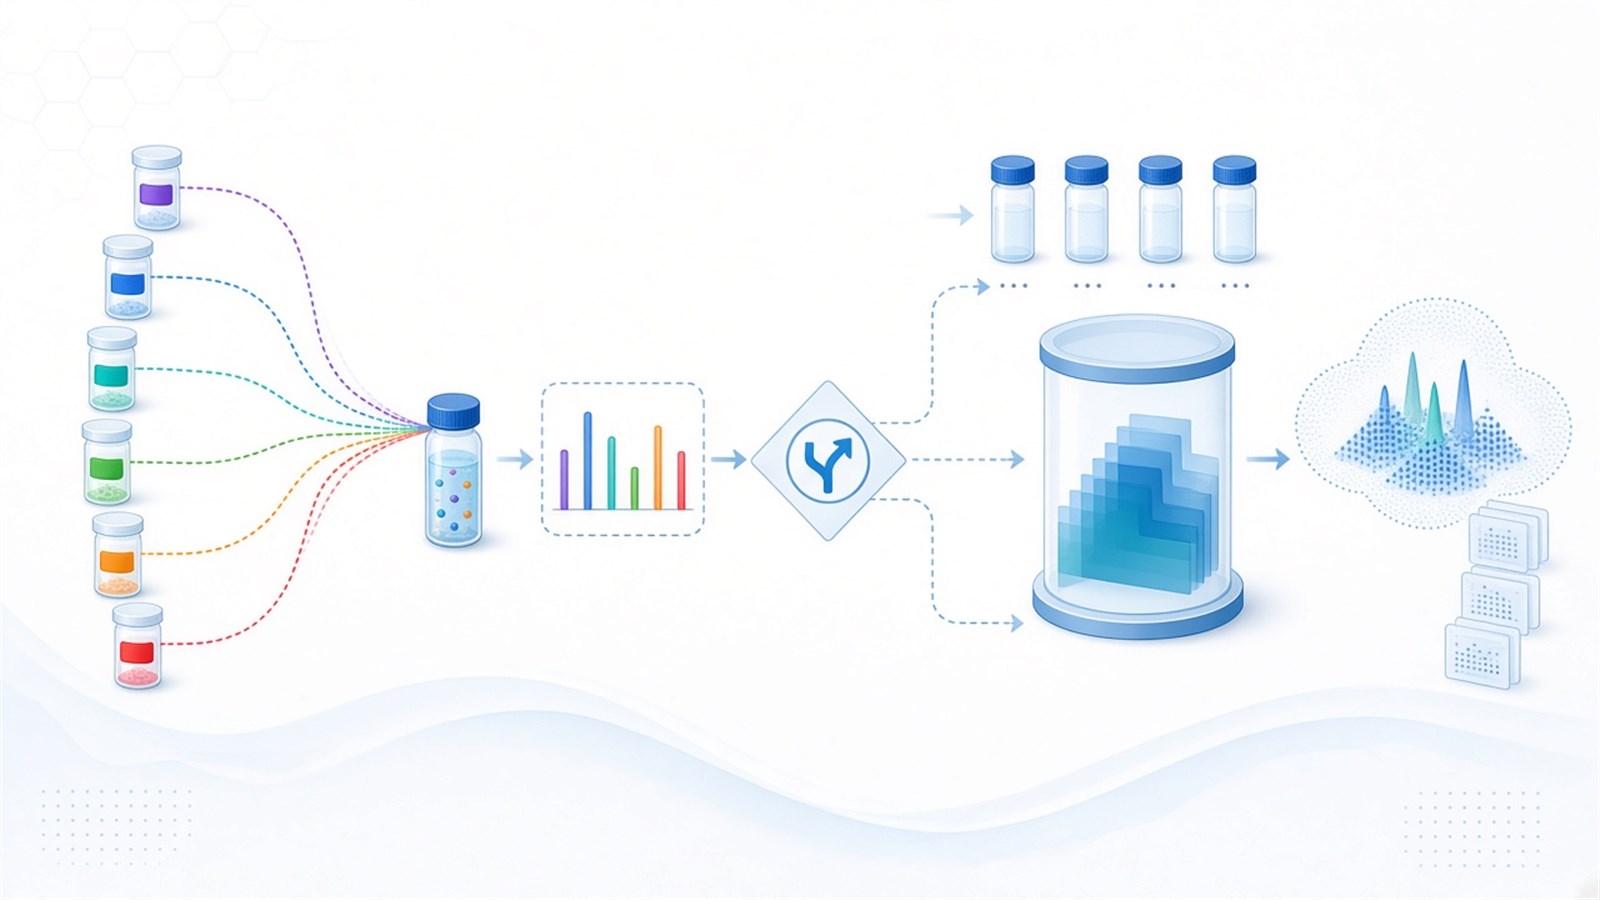
Figure 1. Keep TMT when a locked plex answers the relative claim; branch to SWATH when label-free DIA constraints fit better.
Parameters That Must Stay Locked on Either Route
Layer |
TMT parameter |
SWATH / DIA parameter |
Failure mode if ignored |
|---|---|---|---|
Claim |
Within-plex relative ratio |
Between-injection relative profile |
Cross-wiring reporter and DIA units |
Design |
Channel map and bridges |
Injection order and batch blocks |
Hidden batch as biology |
Prep |
Matched load and labeling completeness |
Matched digest without tags |
Prep drift before acquisition |
Depth |
Fractionation recipe per plex |
Window scheme and gradient |
Unequal depth across arms |
Interference |
MS2 isolation or MS3 reporters |
Library or library-free rules |
Compressed or mis-extracted quantities |
QC |
Bridge and channel QC |
QC pools across injections |
Undetected drift |
Software |
Reporter extraction settings |
DIA extraction settings |
Irreproducible reprocessing |
Hand-off |
Named list for verify |
Named list for verify |
Discovery sold as targeted proof |
Deliverable |
Channel map and method file |
Window/library and method file |
Report rebuilt after acquisition |
Limits follow the same table. High-resolution instruments alone do not fix TMT co-isolation. Complete spectral windows alone do not fix unmatched SWATH prep. Clinical cohort slogans and multi-omics network language need separate study design beyond either table.
How to Choose TMT Versus SWATH After Comprehensive Review
Use multiplex need, cohort growth, claim unit, and follow-up intent together.
Project need |
Route that often fits |
Recheck before committing |
|---|---|---|
Many groups in one mixed LC path |
TMT |
Force SWATH without unlabeled need |
Expanding unlabeled cohort |
SWATH / DIA |
Fit every new sample into one TMT kit |
Strong MS2 ratio-compression risk |
TMT with MS3 / SPS-MS3 plan |
Ignore isolation purity |
Retrospective window re-use |
SWATH / DIA |
Expect TMT reporters to re-open like DIA |
Closed peptide verification |
MRM or PRM after either route |
Treat discovery ratios as targeted assays |
Small contrast, no multiplex pressure |
LFQ or DIA may be simpler |
Buy TMT chemistry without channel need |
Incomplete TMT labeling on pilot |
Fix amines and load first |
Switch to SWATH only to hide labeling failure |
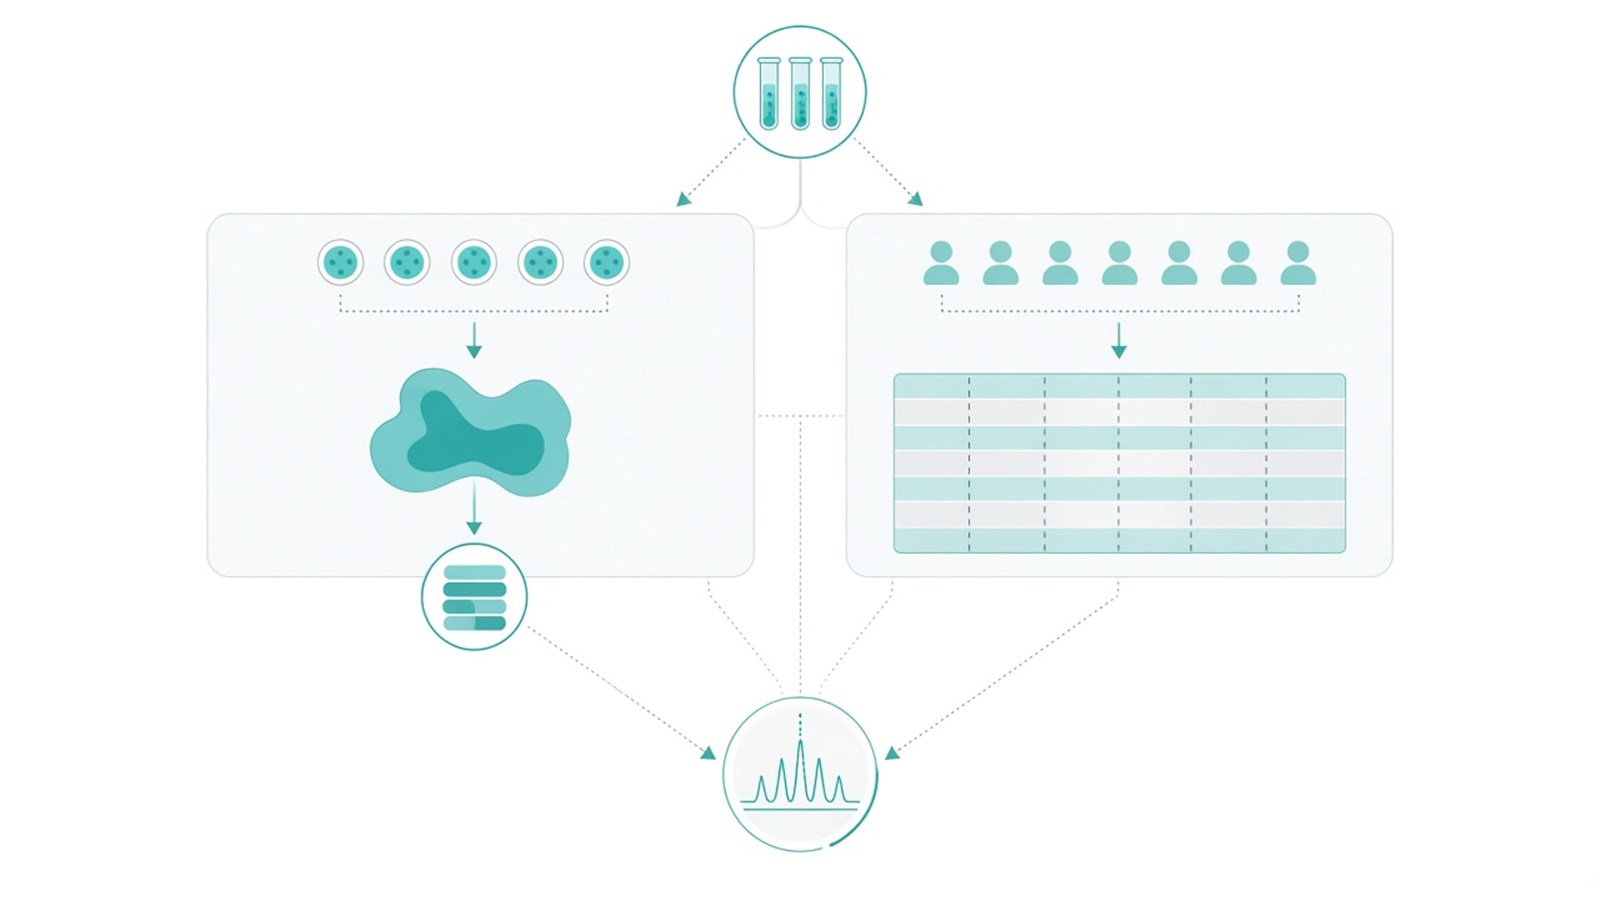
Figure 2. Choose TMT for locked plex relative discovery; choose SWATH when label-free DIA cohort rules fit; keep claim units separate.
Teams with plex maps or SWATH cohort plans can review MtoZ Biolabs TMT or SWATH service entries when matrix, contrast, and route status are declared in the project record.
Frequently Asked Questions
1. Did SWATH replace TMT in quantitative proteomics?
No. TMT and SWATH answer different design constraints. Multiplex labeled mixing and label-free DIA remain complementary routes.
2. Can one study report TMT reporters and SWATH peaks in a single undeclared matrix?
Not without a declared bridging design. Claim units and batch structures differ; mixing them after the fact obscures biology.
3. When should a TMT project still add MS3 instead of switching to SWATH?
When multiplexing remains required and MS2 pilots show ratio compression. Changing to SWATH is a different experiment class, not an MS3 substitute.
4. Is SWATH always better for large clinical cohorts?
Only when unlabeled injections, batch blocks, and extraction rules are locked. Cohort size alone does not choose the route.
5. Should every TMT or SWATH discovery continue into MRM or PRM?
Only when a closed peptide list and verification claim are declared after the discovery contrast.
MtoZ Biolabs can discuss TMT-centered scoping versus SWATH when plex need, cohort growth, and contrast design are documented.
If further evaluation of channel maps, MS3 options, or DIA window plans is needed, project details can be submitted for a feasibility review.
Reference
- A. Thompson, J. Schäfer, K. Kuhn, S. Kienle, J. Schwarz, G. Schmidt, T. Neumann, R. Johnstone, A.K. Mohammed, C. Hamon (2003). Tandem mass tags: a novel quantification strategy for comparative analysis of complex protein mixtures by MS/MS. Anal. Chem., 75, 1895-1904. https://doi.org/10.1021/ac0262560
- L.C. Gillet, P. Navarro, S. Tate, H. Röst, N. Selevsek, L. Reiter, R. Bonner, R. Aebersold (2012). Targeted data extraction of the MS/MS spectra generated by data-independent acquisition: a new concept for consistent and accurate proteome analysis. Mol. Cell. Proteomics, 11, O111.016717. https://doi.org/10.1074/mcp.O111.016717
- L. Ting, R. Rad, S.P. Gygi, W. Haas (2011). MS3 eliminates ratio distortion in isobaric multiplexed quantitative proteomics. Nat. Methods, 8, 937-940. https://doi.org/10.1038/nmeth.1714
- G.C. McAlister, D.P. Nusinow, M.P. Jedrychowski, M. Wühr, E.L. Huttlin, B.K. Erickson, R. Rad, W. Haas, S.P. Gygi (2014). MultiNotch MS3 enables accurate, sensitive, and multiplexed detection of differential expression across cancer cell line proteomes. Anal. Chem., 86, 7150-7158. https://doi.org/10.1021/ac502040v
- M. Bantscheff, S. Lemeer, H.U. Haab, B. Kuster (2012). Quantitative mass spectrometry in proteomics: a critical review. Anal. Bioanal. Chem., 404, 939-965. https://doi.org/10.1007/s00216-012-5892-4
