



A PRM quantitative proteomics workflow from sample to result runs as closed-panel design, matched prep, scheduled high-resolution MS2 acquisition, traceable data processing, and contrast testing on an exported matrix. Operationally, nominate proteotypic peptides from prior discovery, match lysis and digestion across groups, assay protein load, optionally add stable isotope standards for amount designs, build and lock the inclusion list with retention-time windows, acquire PRM on each sample, extract and review fragment chromatograms, normalize with QC pools, then statistics on predefined arms only (Peterson et al. 2012; Lange et al. 2008). Default output is relative peptide signal unless SIS and calibration are declared. Reserve aliquots before digestion when confirmation may expand.
Researchers who already have a closed peptide panel or a validation objective can also review the MtoZ Biolabs Parallel Reaction Monitoring (PRM) Service and Targeted Proteomics Service for project-specific feasibility and workflow planning.
End-to-End Workflow: Sample Through Reported Matrix
Stage |
Key operations |
Document before scaling |
|---|---|---|
Design |
Contrast arms, replicates, relative or absolute claim |
Frozen hypothesis list |
Panel |
Proteotypic peptides, charges, modifications |
Target list version |
Prep |
Extract, reduce, alkylate, digest, desalt |
Matched recipe per arm |
Load QC |
Protein assay; consistent digest input |
mg loaded per sample |
Method |
Schedule precursors, RT windows, cycle time |
Inclusion table locked |
Optional SIS |
Spike timing and curve plan for amount units |
Matrix-matched calibrators |
Acquisition |
HR MS2 PRM per sample; QC interleave |
Raw files tied to IDs |
Processing |
XIC extract, peak review, normalization |
Software version locked |
Results |
Export matrix; predefined stats |
No post hoc arm changes |
- Close the peptide panel after discovery evidence; PRM cannot quantify proteins absent from the schedule.
- Prepare samples with the same digestion chemistry for every contrast arm.
- Develop scheduling on pilot injections before full cohort acquisition.
- Process raw files with documented integration rules; revisit peaks when interference appears.
- Interpret group differences only inside the declared claim; pathway tools are secondary.
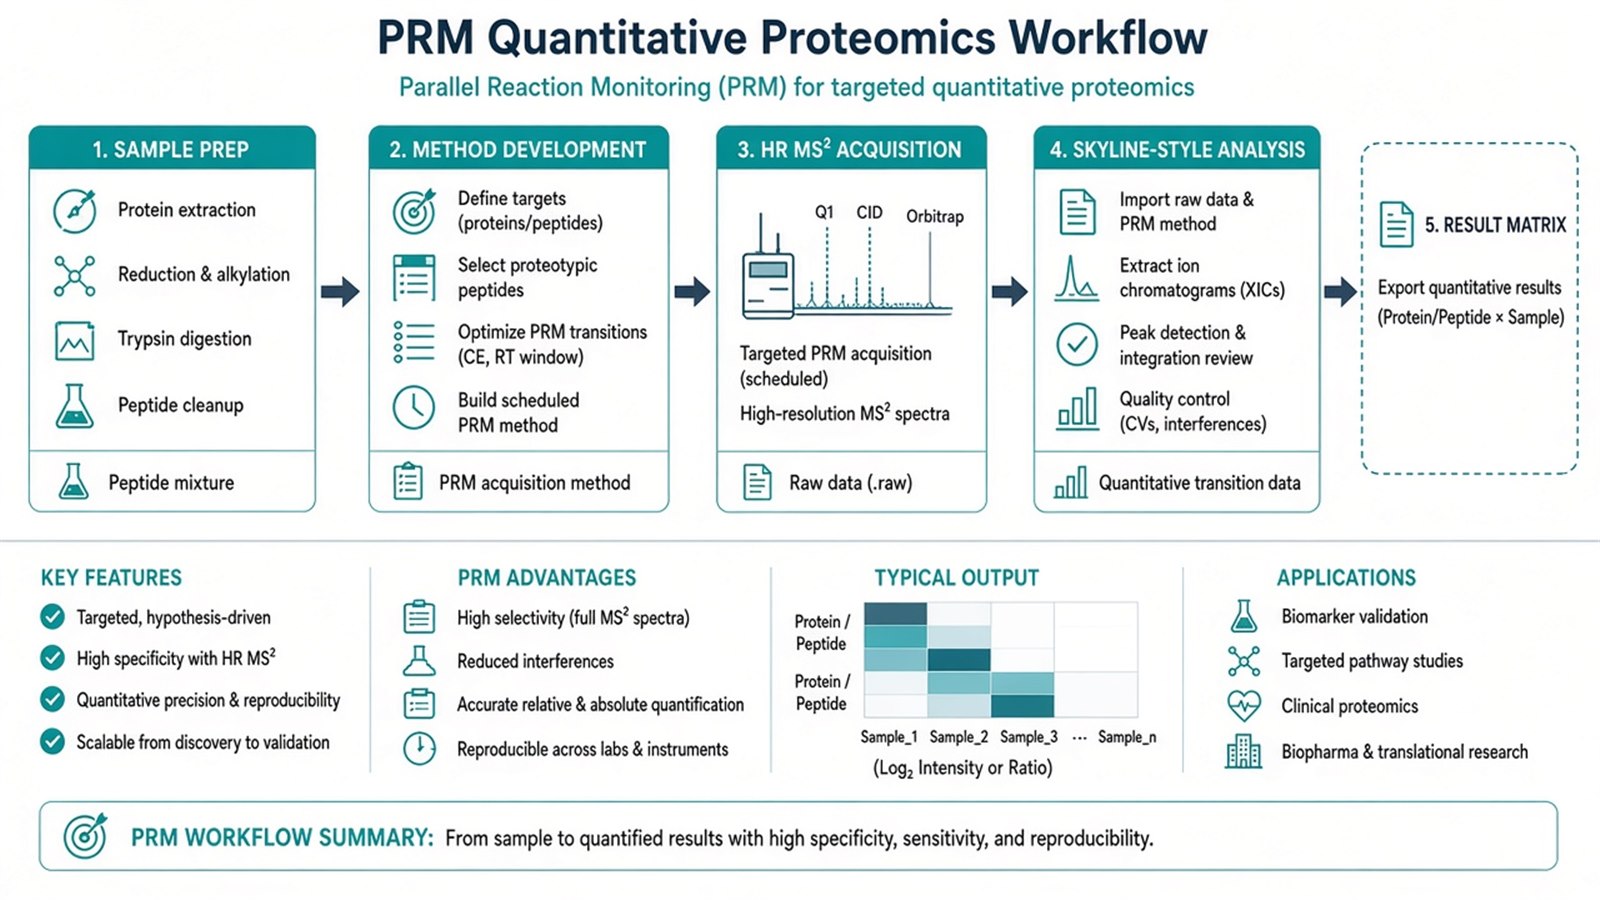
Figure 1. Sample-to-result PRM workflow links bench prep, targeted acquisition, and traceable analysis before biological interpretation.
Calling enrichment analysis part of the PRM result without matrix QC overstates what the targeted run alone defends.
For processing detail, see PRM Proteomics Data Processing Workflow. For analysis tools, see Tools for PRM Proteomics Data Analysis. For validation design, see How to Plan a Targeted Proteomics Validation Study.
Sample Prep and Acquisition Checkpoints
PRM specificity at the instrument still depends on prep parity and realistic scheduling.
Checkpoint |
Pass criteria |
Common failure |
|---|---|---|
Extraction |
Comparable yield across arms |
Arm-specific lysis bias |
Digestion |
Stable peptide recovery |
Overdigestion or detergent carryover |
Load |
Mass-based digest input |
Volume loading hides yield |
Panel |
Unique surrogates per protein |
Shared peptides across isoforms |
Schedule |
Enough points across peaks |
Over-multiplexed inclusion list |
SIS |
Spike recorded before critical loss |
Late spike misses upstream loss |
Raw QC |
Stable RT and mass accuracy |
Batch day tracks group labels |
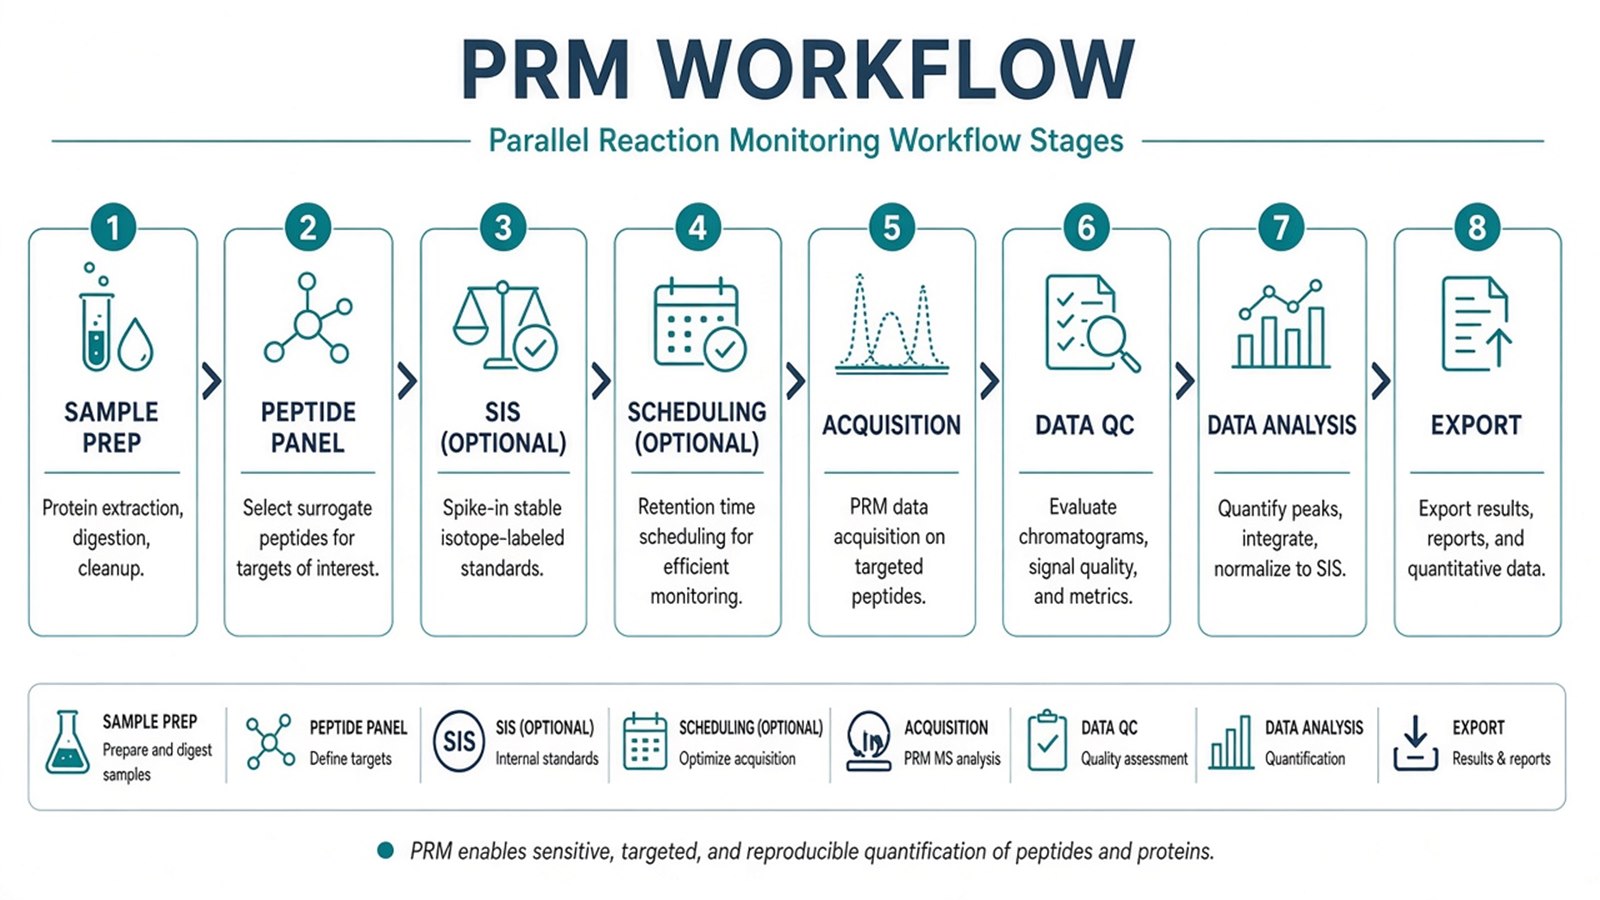
Figure 2. Sample-to-result reliability is set by prep, panel closure, and scheduling before full cohort acquisition.
For mechanism context, see Mechanism of MRM/PRM in Protein Quantification. For discovery handoff, see Application of DIA-PRM Proteomics and Detection of Proteins Using Parallel Reaction Monitoring.
Judgment Criteria: When a Full PRM Workflow Fits
Study need |
Sample-to-result PRM fits |
Reassess |
|---|---|---|
Inventory |
Peptides nominated and schedulable |
Open discovery still expanding |
Claim |
Relative confirmation or SIS absolute on panel |
Diagnostic cutoff without validation |
Matrix |
Complex digest needs HR MS2 review |
Mature MRM transitions suffice |
Material |
Enough input for pilots and QC |
Single aliquot with no reserve |
Timeline |
Time for method lock before cohort |
Rush acquisition before panel QC |
Prefer PRM when a closed list needs high-resolution fragment evidence from sample to matrix. Prefer MRM when mature transitions and triple-quadrupole multiplex dominate. Prefer DIA or LFQ while candidates remain unknown. Prefer AQUA-style absolute designs when concentration units and curves are planned on the same shortlist.
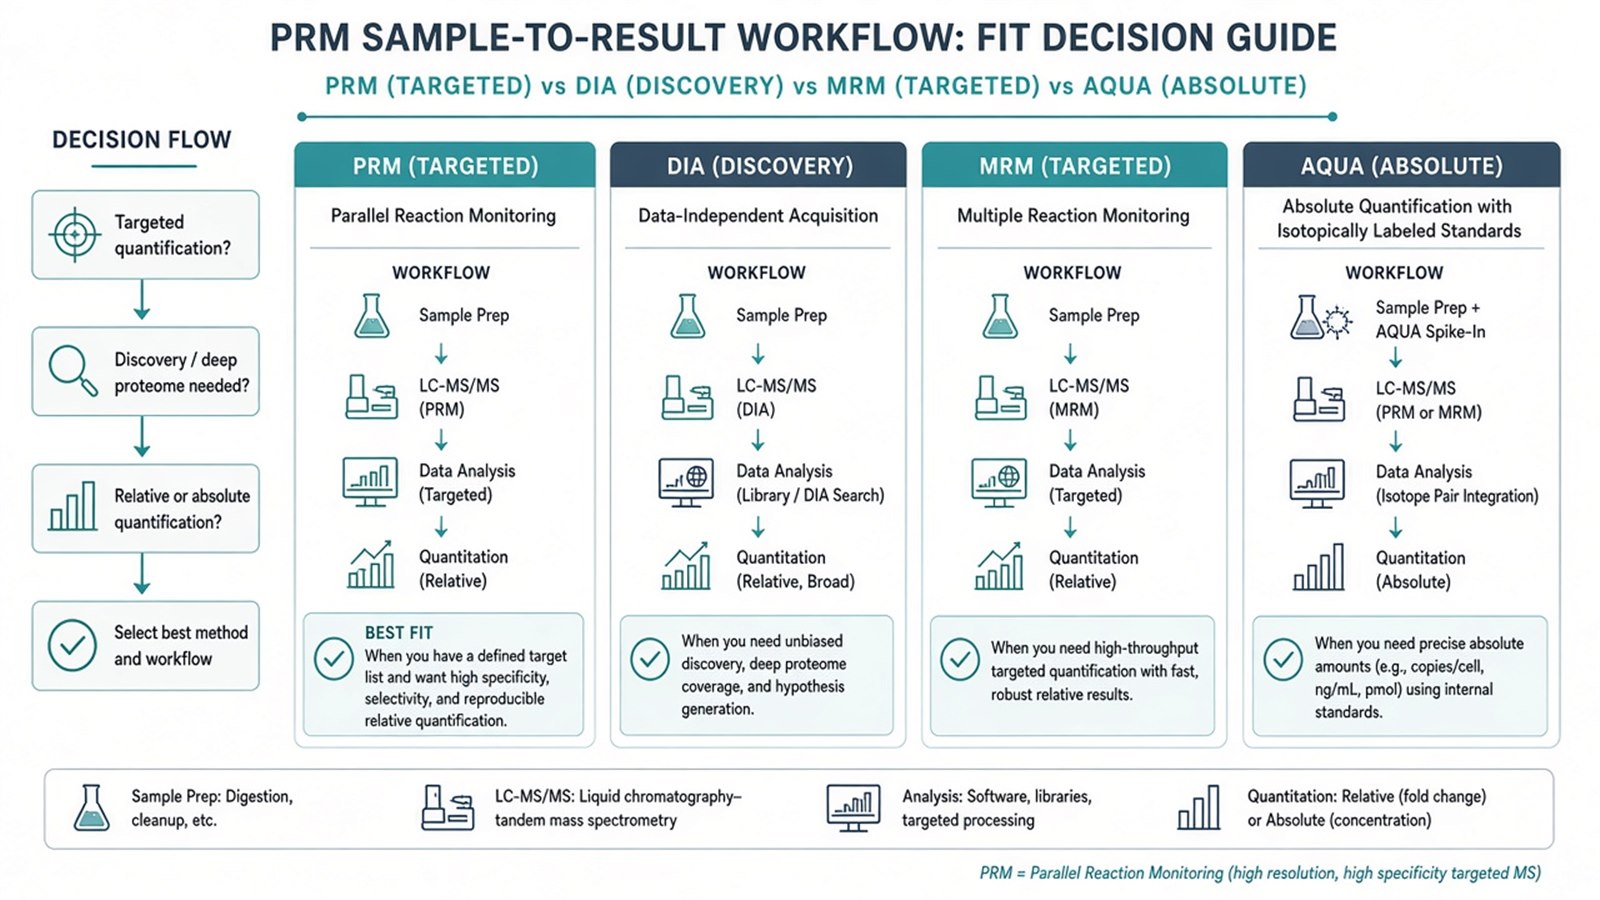
Figure 3. Use the full PRM workflow when prep, panel, and scheduling can be locked; otherwise stay on discovery or alternate targeted routes.
For targeted quantification framing, see Parallel Reaction Monitoring for Targeted Quantification. For advantage trade-offs, see Advantages and Disadvantages of Parallel Reaction Monitoring. For absolute steps, see Absolute Quantification in Targeted Proteomics.
Limits and Application Boundaries
A detailed workflow does not make PRM an open proteome survey. Co-isolation can still distort fragments if peaks are integrated without review. Relative matrices are not clinical cutoffs. Protein inference depends on surrogate peptides. Pathway enrichment reflects the quantified panel background, not whole-proteome mechanism proof.
Typical sample-to-result applications:
- Biomarker-candidate verification after discovery.
- Pharmacodynamic panels across treatment arms on fixed peptides.
- Biopharmaceutical peptide monitoring with locked methods.
- Multi-batch bridging when QC pools and anchor peptides are designed in advance.
- SIS-backed amount readouts when calibration is part of the same workflow record.
Diagnostic deployment sits outside ordinary research PRM reporting unless separately validated.
Once panel, prep, schedule, processing version, and contrasts are locked, the project can deliver a defensible PRM result matrix. Researchers can review the MtoZ Biolabs Parallel Reaction Monitoring (PRM) Service for sample intake and workflow planning.
Frequently Asked Questions
1. When should the peptide panel close in a PRM workflow?
Before full cohort acquisition, after pilot scheduling shows usable transitions and retention times.
2. Is SIS mandatory in PRM?
No. SIS is required only when the claim includes amount units with calibration, not for default relative confirmation.
3. Can PRM start without prior discovery?
Only when peptides are already justified by literature or pilot data; otherwise discovery or DIA usually comes first.
4. Where does Skyline fit in sample to result?
At processing: target list management, XIC review, normalization, and export before external statistics (MacLean et al. 2010).
5. What is outside a standard PRM result deliverable?
Diagnostic cutoffs, companion-diagnostic claims, and whole-proteome discovery without a new acquisition design.
Reference
- A.C. Peterson, J.D. Russell, D.J. Bailey, M.S. Westphall, J.J. Coon (2012). Parallel reaction monitoring for high resolution and high mass accuracy quantitative, targeted proteomics. Mol. Cell. Proteomics, 11, 1475-1488. https://doi.org/10.1074/mcp.O112.020131
- V. Lange, E. Picotti, B. Domon, R. Aebersold (2008). Selected reaction monitoring for quantitative proteomics: a tutorial. Mol. Syst. Biol., 4, 222. https://doi.org/10.1038/msb.2008.61
- B. MacLean, D.M. Tomazela, N. Shulman, et al. (2010). Skyline: an open source document editor for creating and analyzing targeted proteomics experiments. Bioinformatics, 26, 966-968. https://doi.org/10.1093/bioinformatics/btq054
