



A Co-IP protocol, or co-immunoprecipitation protocol, asks which proteins remain with an endogenous bait after antibody capture, Input and IgG contrast, and either targeted detection or LC-MS/MS identification. Designing that protocol means locking an IP-competent antibody, matched control arms, and a wash-elution recipe that can show bait enrichment before any partner is ranked. The design fits cells or tissues when native complexes can be recovered and the claim is co-recovery, not binary direct binding (Hein et al. 2015). Prefer this route when antibody performance can be shown in a pilot. Reassess when the antibody is Western-only, Input and IgG are omitted, or enrichment is too weak for a credible list.
Researchers who already have a named bait, a planned Co-IP protocol, and a written interaction claim can also review the MtoZ Biolabs IP-Mass Spectrometry Service and Protein-Protein Interaction Analysis for project-specific protocol feasibility.
Design the Co-IP Protocol Around the Written Claim
A Co-IP protocol is efficient when every step can support the sentence the study will write. It is not efficient because a lysis buffer or wash count was copied from another bait. Write the claim first: confirm a named pair, discover unknown partners, or compare the same bait across conditions.
Confirmation usually keeps a tighter wash and a targeted readout once bait recovery is shown. Discovery usually keeps milder native lysis so associated proteins can remain, then ranks names against IgG. Condition comparison needs the same antibody, lysis, wash, and elution on every arm so relative intensity of the same protein can be compared. A transferred TMT or label-free recipe does not replace that matched design.
The protocol can end as Co-IP western blot or as IP-MS. Both share antibody capture. They diverge at whether the eluate is read for named prey or identified by LC-MS/MS. That readout choice is compared in IP-MS vs Co-IP Western Blot: Which Method Fits Your Study?. Capture-to-MS layers after the design is locked are covered in IP-MS Protocol: From Antibody Capture to LC-MS/MS.
Native lysis is a design constraint, not a brand of buffer. Milder detergents often retain cytoplasmic or membrane-associated assemblies. Harsher native mixes can keep stable complexes and raise background. Denaturants that unfold the bait belong outside a Co-IP protocol that still claims associated proteins. Pilot the lysis that recovers the bait with acceptable partner retention, rather than the harshest mix that looks clean on blot.
A Co-IP protocol is designed around a written co-recovery claim, an IP-competent antibody, and matched Input and IgG arms. It is not a copied wash list.
Build Antibody Specificity Into Capture and Wash
Antibody specificity belongs in the protocol, not in a later note. The capture reagent must recover the intended native bait from the planned lysate without a dominant off-target that then looks like a partner. Application-specific validation remains necessary because Western performance does not establish IP performance (Uhlen et al. 2016). How to choose that reagent is covered in How to Choose the Right Antibody for Co-IP Assays?.
In the protocol itself, record clone, host species, isotype, lot, and the bead chemistry used to collect the immune complex. Protein A, protein G, or mixed A/G pairing is part of capture design. A mismatch can look like a failed Co-IP even when the antibody can bind the bait in solution. Covalent coupling of the antibody to beads can lower heavy-chain and light-chain carryover in a later digest or blot. Coupling does not prove that remaining names are partners.
Wash stringency is the main specificity-versus-retention trade-off after the antibody is chosen. Gentle washes keep weak or transient associations and raise IgG-like background. High-salt or detergent-rich washes can clear sticky binders and can also strip the associations the claim needed. Do not set a universal wash count. Titrate wash chemistry against bait recovery in the bait IP and against the same proteins in the matched IgG arm.
Input and IgG Are Designed Arms, Not Afterthoughts
Input and IgG must be written into the Co-IP protocol before the first incubation. They are the tests that make antibody specificity and enrichment efficiency readable.
Input is taken before capture. It shows whether the bait was present and extractable, and it provides a baseline for enrichment. If Input lacks the bait, a missing IP signal is an expression or extraction problem. It is not a finished interaction result.
IgG or bead controls must use the same lysate batch, lysis chemistry, incubation time, wash stringency, bead type, and elution as the bait IP. The isotype arm estimates handle-driven background. It cannot replace Input. Empty-tag controls from AP-MS do not replace IgG in an antibody Co-IP protocol. Asymmetric wash or elution between bait IP and IgG makes specificity look better or worse than it is.
Protocol arm |
Design job |
What it cannot do |
|---|---|---|
Antibody plus bead pairing |
Recover the native bait without a dominant off-target |
Convert a Western-only reagent into IP-grade capture |
Input |
Show starting bait and a baseline for enrichment |
Prove the IP antibody works |
IgG or bead |
Estimate handle-driven background under the same recipe |
Replace Input or prove a remaining name is a partner |
Wash and elution |
Set retention versus background, then release material for blot or MS |
Rescue a failed antibody |
Replicates |
Stabilize the same recipe across biological arms |
Merge unlike lysis or wash chemistries |
A practical design sequence is:
- Write the co-recovery claim and the intended readout.
- Qualify native bait recovery with the chosen antibody versus Input.
- Process a matched IgG or bead arm with the same recipe.
- Titrate wash and elution against bait enrichment, not against a clean-looking blot alone.
- Only then rank co-enriched names or bands.
Sample-prep parity after those arms exist is covered in How to Prepare Immunoprecipitation Samples for Mass Spectrometry.
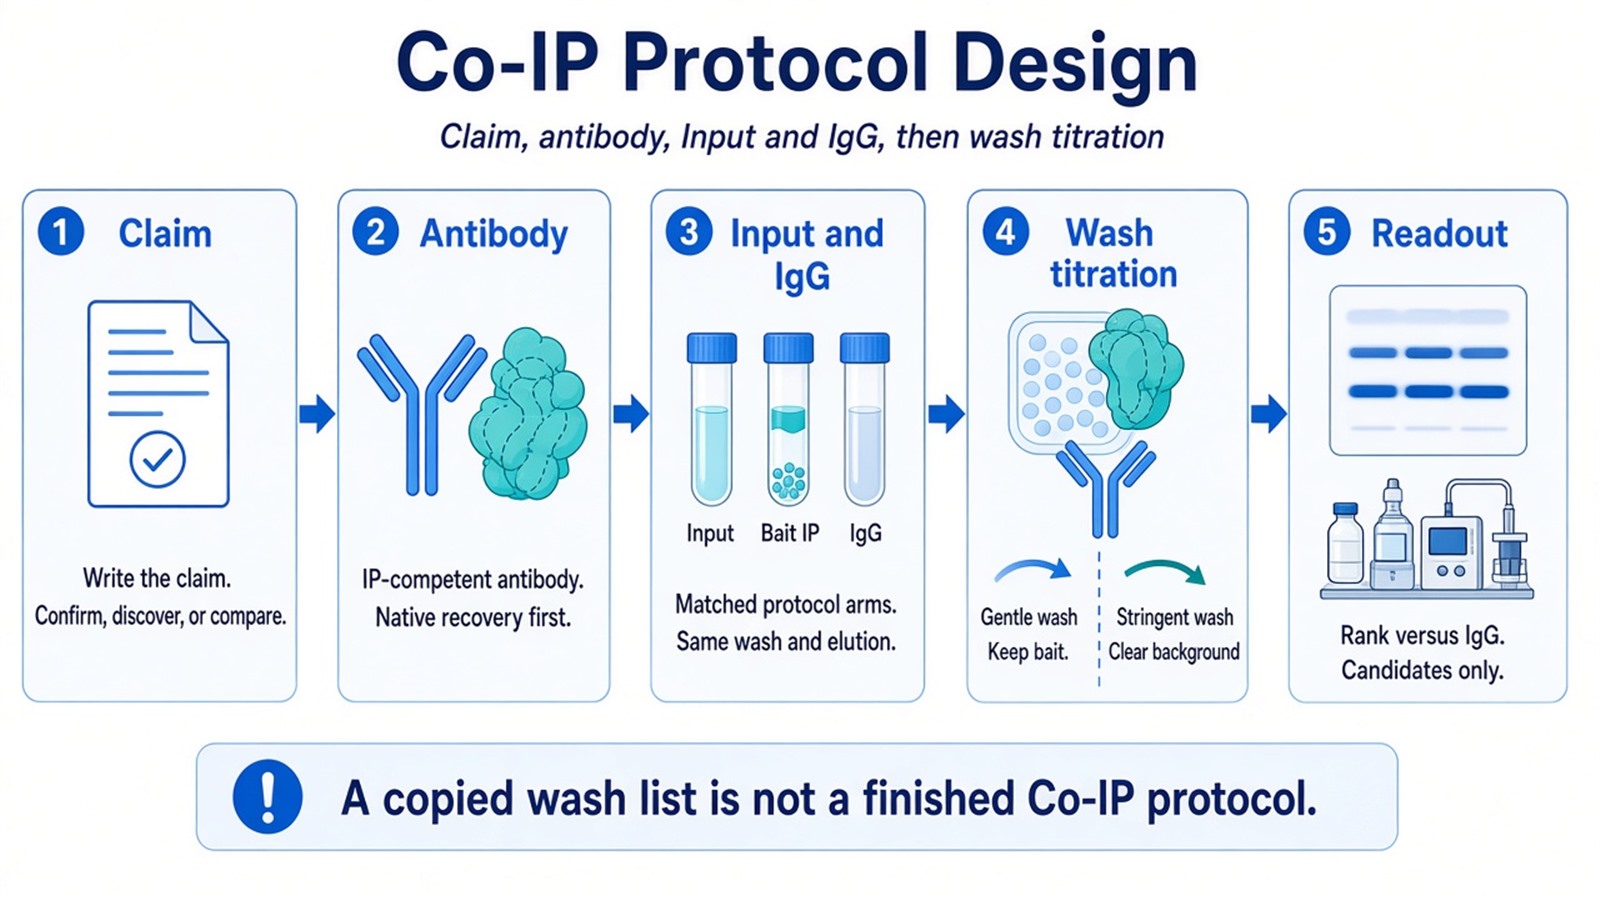
Figure 1. Protocol design starts with the claim and the antibody. Wash counts are titrated after Input and IgG arms exist.
Enrichment Efficiency and Mass Spectrometry Credibility
Enrichment efficiency is bait-first. If the bait is weak versus Input, a long identification table is not a stronger interactome, and a stronger prey band is not a stronger protocol. Relative intensity can compare the same protein across Input, IgG, and bait IP. It does not rank unrelated proteins as tighter binders.
For a blot ending, heavy-chain and light-chain bands can hide a prey of similar mass. Chain-blind detection can reduce that readout problem. It does not repair upstream capture. For an IP-MS ending, antibody chains and bead proteins can dominate peptide sampling when antibody input is high relative to bait recovery. Identification depth in a Co-IP-style eluate is typically hundreds to about a thousand proteins, depending on the sample. Depth without bait enrichment is background.
Identification of a protein in the eluate is not a confidence call. Ranking requires bait recovery, Input-aware enrichment, and a declared contrast against IgG. Probabilistic scoring such as SAINT uses bait versus control counts to assign a probability that a prey is specifically co-purified (Choi et al. 2011). A contaminant catalog such as the CRAPome is a prior on control frequency, not a substitute for the current Input and IgG pair (Mellacheruvu et al. 2013). Neither score repairs a protocol that omitted those arms.
After those gates, a protein-level value can support a relative statement: protein B is enriched with bait A versus matched IgG inside this Co-IP protocol. Peptide support, antibody clone, wash chemistry, and elution route belong next to that statement. Pathway maps do not repair a missing IgG arm.
If bait recovery fails after the protocol is run, diagnose before ranking prey, as in Why Was the Target Protein Not Detected by IP-MS?.
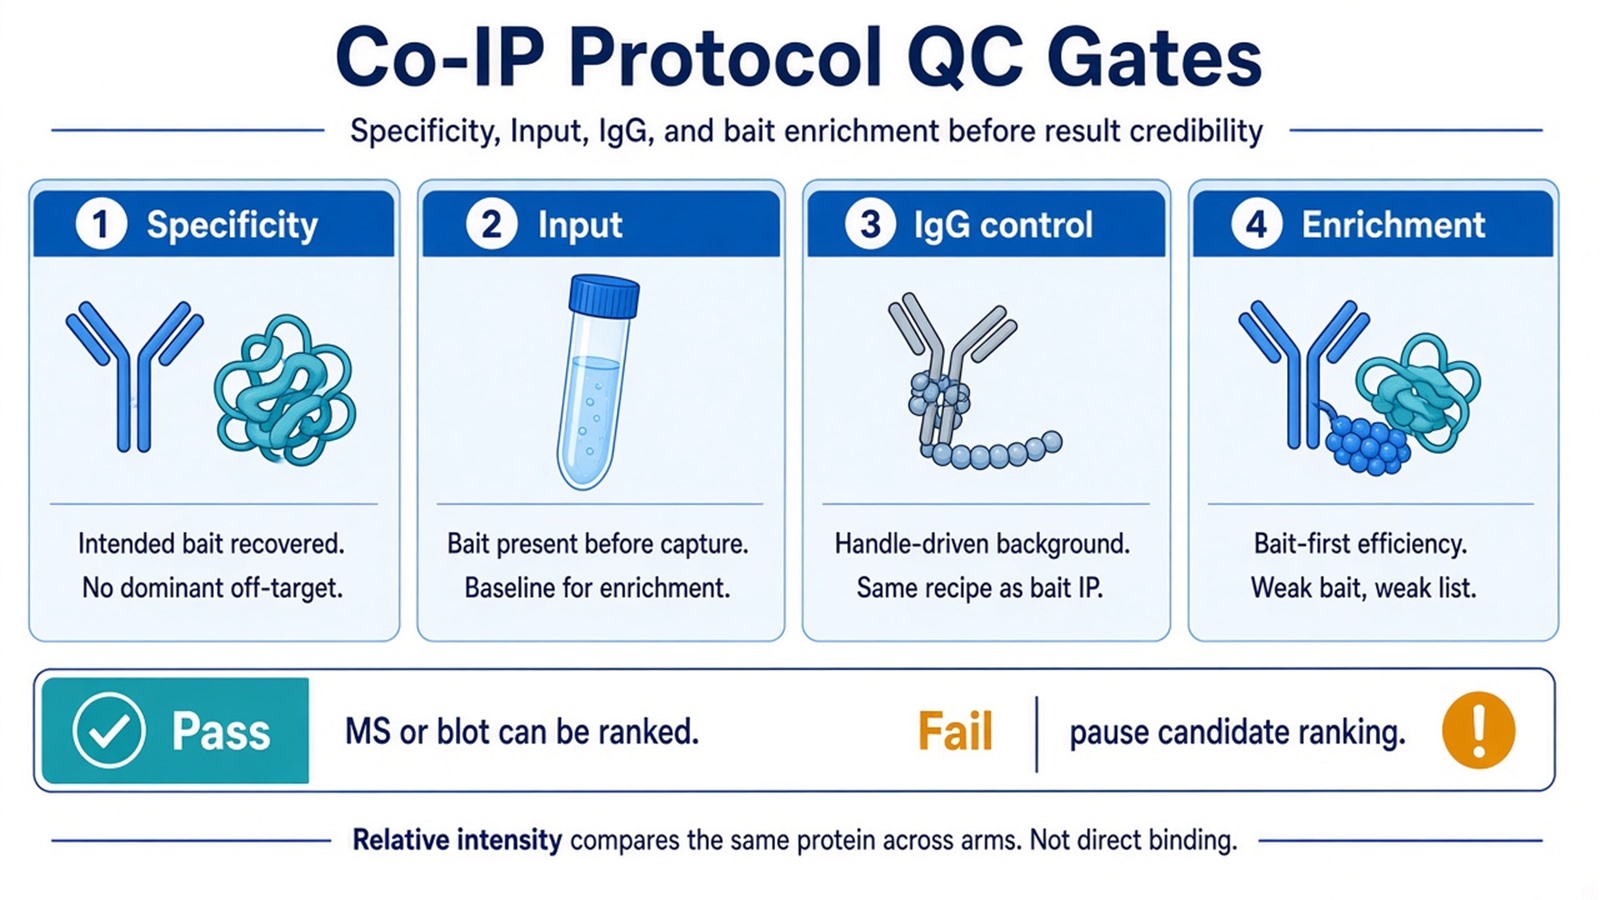
Figure 2. Mass spectrometry credibility is a protocol outcome. It starts after bait enrichment is shown versus Input and IgG.
Common Biases and Result Boundaries
The common design bias is copying a published Co-IP protocol and treating the first eluate as an interactome. A recipe that worked for one bait can fail when the antibody, lysis, or complex stability changes. Another bias is adding more wash steps to chase a clean blot, then concluding that missing partners mean no interaction. Under-washing creates the opposite error: bait IP and IgG look similar, so subtraction makes every name look specific.
Do not write a successful bait IP as proof that every co-enriched protein contacts the bait. Do not write a failed Co-IP protocol as proof that no interaction exists in the cell. Absence of bait recovery can mean the wrong antibody, masked epitope, incompatible bead chemistry, over-washing, or a bait that is not expressed in the tested state.
Revise antibody, lysis, or wash when Input is positive and IP recovery is not. Prefer AP-MS or an immobilized recombinant bait when no IP-competent antibody exists and tagging or purified bait is acceptable. Those switches are compared in AP-MS vs IP-MS: How to Choose the Right Interactome Workflow. Screening-level method maps are covered in Measuring Protein-Protein Interactions. Mixed enrichment-MS projects can be reviewed under the MS-Based Protein-Protein Interaction Analysis Service.
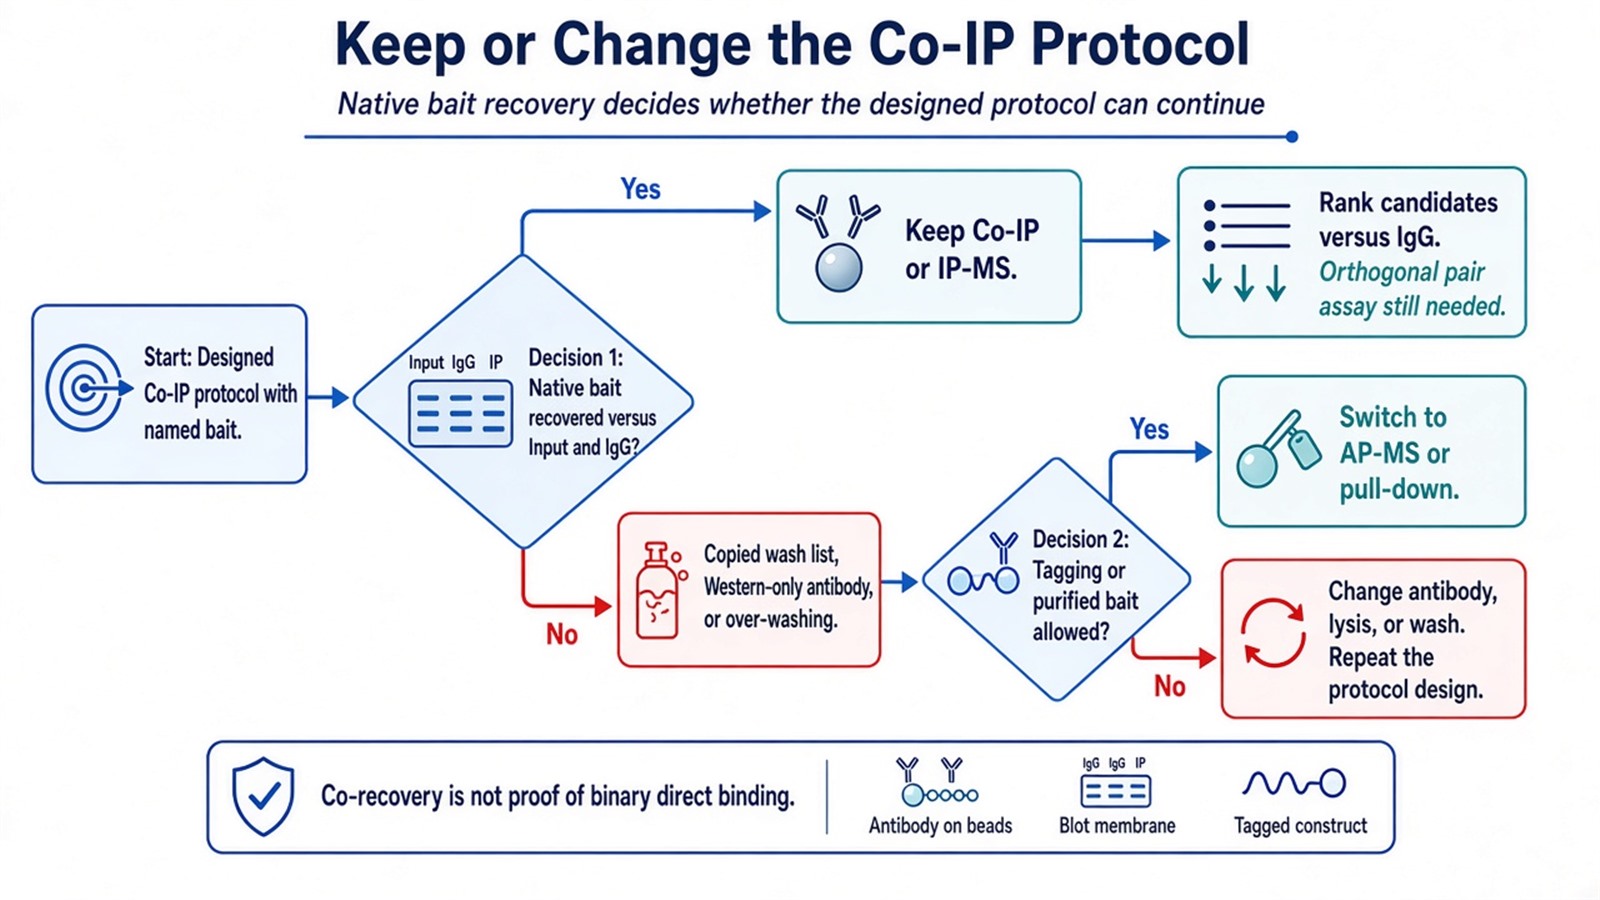
Figure 3. Stay with the Co-IP protocol only while native bait recovery is documented. A copied wash list is not a completed design.
A designed Co-IP protocol supports candidate co-recovery after antibody, Input, IgG, and bait enrichment are shown. It does not certify a binary interactome.
Once the claim, antibody status, Input and IgG plan, enrichment checks, and the intended blot or IP-MS readout are defined, the project can be scoped as a Co-IP or IP-MS study. Researchers can review the MtoZ Biolabs IP-Mass Spectrometry Service for sample evaluation and protocol planning.
Frequently Asked Questions
1. Can I transfer a published Co-IP protocol to my bait?
Only after native bait recovery is re-shown versus Input with your antibody and lysis. A published wash list is a starting draft, not a finished design.
2. How many washes should a Co-IP protocol use?
Titrate wash chemistry against bait recovery in the bait IP and against the same proteins in matched IgG. A copied 3-to-5 wash rule is only a starting draft.
3. Can IgG replace Input in a Co-IP protocol?
No. Input tests whether the bait was present before capture. IgG tests handle-driven background. The two arms answer different questions.
4. When does a Co-IP protocol become an IP-MS protocol?
When the same designed arms are digested and identified by LC-MS/MS, and candidates are ranked against matched IgG rather than called by presence alone.
5. Does coupling the antibody to beads replace an IgG control?
No. Coupling can lower antibody-chain carryover. It does not estimate lysate proteins that bind the handle or the beads.
Reference
- M. Uhlen, A. Bandrowski, S. Carr, A. Edwards, J. Ellenberg, E. Lundberg, D. L. Rimm, H. Rodriguez, T. Hiltke, M. Snyder, T. Yamamoto (2016). A proposal for validation of antibodies. Nat. Methods, 13, 823-827. https://doi.org/10.1038/nmeth.3995
- M.Y. Hein, N.C. Hubner, I. Poser, J. Cox, N. Nagaraj, Y. Toyoda, I.A. Gak, I. Weisswange, J. Mansfeld, F. Buchholz, et al. (2015). A human interactome in three quantitative dimensions organized by stoichiometries and abundances. Cell, 163, 712-723. https://doi.org/10.1016/j.cell.2015.09.053
- H. Choi, B. Larsen, Z.Y. Lin, A. Breitkreutz, D. Mellacheruvu, D. Fermin, Z.S. Qin, M. Tyers, A.C. Gingras, A.I. Nesvizhskii (2011). SAINT: probabilistic scoring of affinity purification-mass spectrometry data. Nat. Methods, 8, 70-73. https://doi.org/10.1038/nmeth.1541
- D. Mellacheruvu, Z. Wright, A.L. Couzens, J.P. Lambert, N.A. St-Denis, T. Li, Y.V. Miteva, S. Hauri, M.E. Sardiu, T.Y. Low, et al. (2013). The CRAPome: a contaminant repository for affinity purification-mass spectrometry data. Nat. Methods, 10, 730-736. https://doi.org/10.1038/nmeth.2557
