



Applications of liquid chromatography-tandem mass spectrometry (LC-MS/MS) in label-free quantitative proteomics (LFQ) are study designs that compare the same protein across independently injected, unlabeled digests. LC separates peptides. MS1 chromatographic peak area usually carries the abundance value. MS2 fragment spectra assign peptide identity. The application is the scientific contrast, not a disease chapter and not a service catalog. An LFQ LC-MS/MS application can support relative change of the same protein inside one chromatographic and processing plan. It cannot report a diagnostic cutoff, a calibrated tissue concentration, or a copy-number ranking across different proteins. Search titles such as application of label-free quantitative proteomics point to the same unlabeled comparison, whether or not LC-MS/MS appears in the heading.
Researchers who already have defined samples, comparison groups, or a specific project objective can also review the MtoZ Biolabs Label-Free Quantitative Proteomics Service for project-specific feasibility and analysis planning.
What an LFQ LC-MS/MS Application Can Claim
The usable application starts from a sentence that has a comparison: treatment versus control, genotype, dose, time point, or tissue state. Proteins are digested, most often with trypsin. Peptide ions are the operational measurement (Bantscheff et al. 2007). After alignment and normalization, a higher protein-level value can support a relative abundance change. Pathway maps, drug-target lists, and biomarker narratives are interpretations of that table. They are not additional LC-MS/MS measurements.
Colorimetric assays such as bicinchoninic acid (BCA) match digest load. They do not identify which proteins changed. Enzyme-linked immunosorbent assay (ELISA) and Western blot quantify selected proteins when antibodies are adequate. They do not replace an untargeted LFQ comparison. Isobaric labeling belongs when many conditions must share one pooled injection. A short peptide panel with a calibration curve belongs to targeted LC-MS/MS.
Researchers can refer to What Is Label-Free Quantification in Proteomics? for how precursor peak area becomes a protein-level matrix.
An LFQ LC-MS/MS application supports relative change of one protein inside one study, not a clinical or agricultural certificate.
Applications That Match Independent LC-MS/MS Runs
Treatment, genotype, and tissue-state contrasts
Discovery comparisons of treated versus untreated cells, knockout versus parental lines, or diseased versus adjacent tissue are the most common LFQ LC-MS/MS applications. The design works when groups are randomized across run order and the chromatographic regime stays one plan. Differential proteins remain candidates. A protein that changes with tumor grade is not, by that fact, a diagnostic marker. A protein that changes after a compound is added is not proof of a drug target.
Time-course and later enrollment
Unlabeled LC-MS/MS fits contrasts that cannot be frozen inside one labeling kit: serial collections, samples that arrive over a planned window, or cohorts that may still grow. Each digest stays its own run. Later files can join only when the analytical regime remains compatible. A column change or a long interruption is a new block. Ordinary normalization does not insert that block into the original matrix.
Matrices that do not enter labeling cleanly
Clinical tissues, biofluids, archival material, and some extracellular-vesicle preparations are frequent LFQ applications because metabolic labeling is unavailable and chemical labeling may consume material or fail in difficult buffers. Compatibility still depends on extraction, contaminants, and a searchable protein database. Naming plasma, exosomes, or cerebrospinal fluid does not relax run-order control or missing-value review.
Plant, food, or environmental matrices can be measured when those same conditions hold. Those fields are sample sources. They are not separate LFQ methods, and they do not authorize allergen release testing or regulatory environmental claims from an ordinary discovery table.
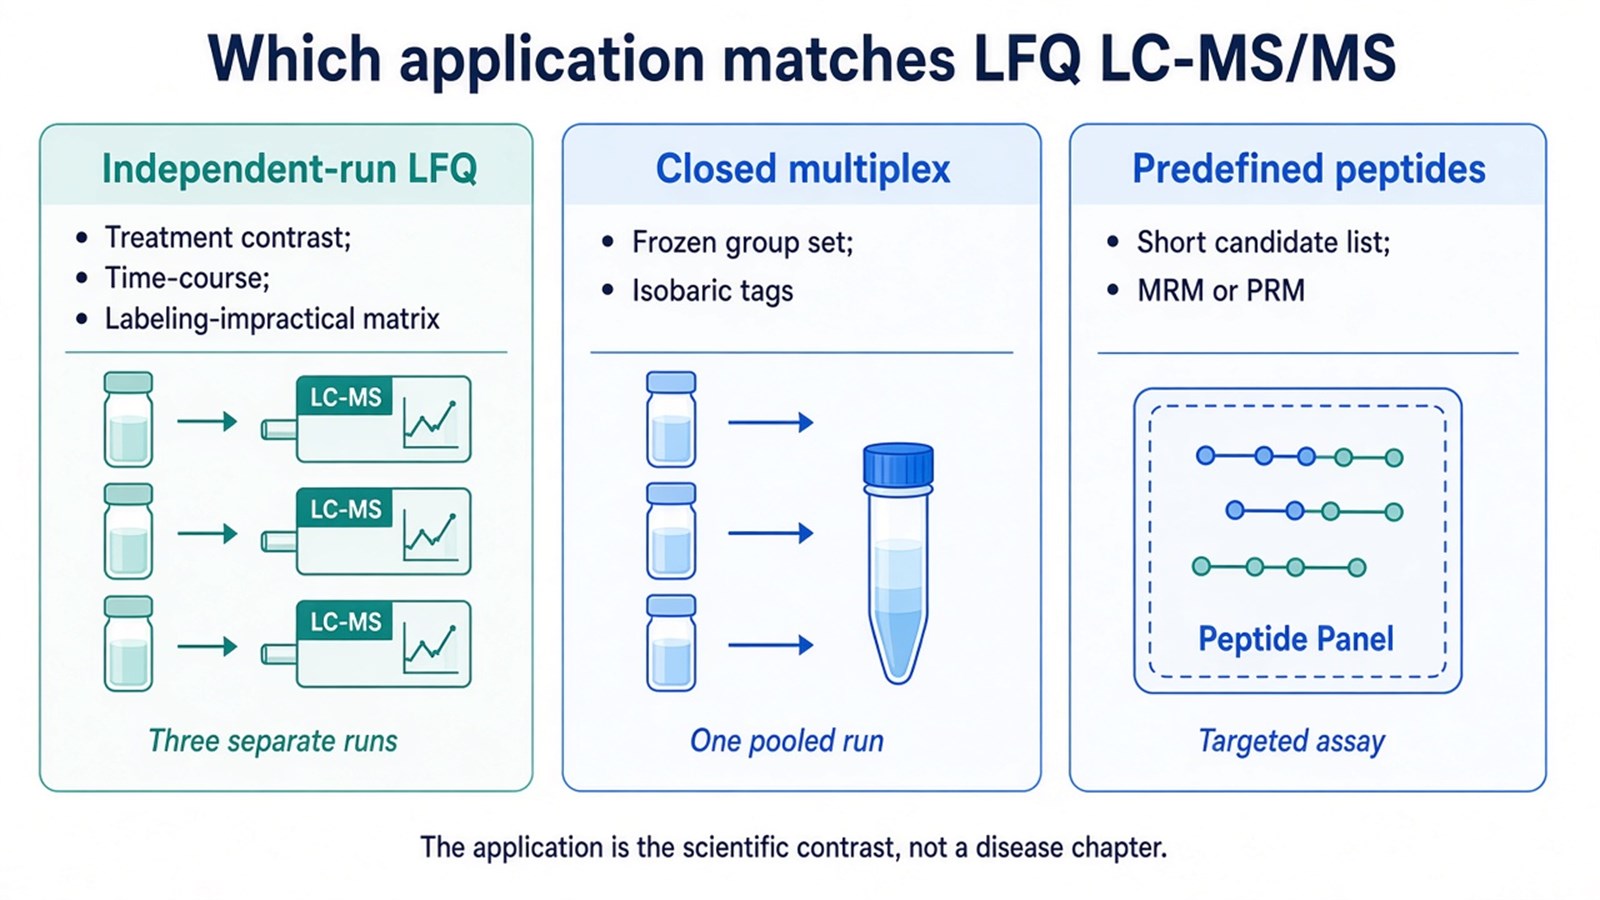
Figure 1. LFQ LC-MS/MS applications are unlabeled independent-run contrasts; a closed multiplex or a predefined peptide amount is a different application.
Applications That Need a Different Assay
Reassess unlabeled LC-MS/MS when the application requires an amount rather than a ratio, a predefined-target confirmation, or a same-run comparison that independent chromatography cannot support. Tandem mass tags (TMT) trade labeling chemistry for reporter-ion ratios inside one mixed run. Multiple reaction monitoring (MRM) or parallel reaction monitoring (PRM) follows a short candidate list. Stable isotope labeling by amino acids in cell culture (SILAC) requires a system that incorporates heavy amino acids.
Data-dependent acquisition (DDA) and data-independent acquisition (DIA) change how complete an unlabeled application matrix is likely to be. They do not change the scientific claim. DIA can reduce missing identifications in larger cohorts (Gillet et al. 2012). DIA is not a new application domain, and it is not defined by the absence of a label.
When sample number, multiplexing, or later enrollment makes the application less straightforward, researchers can refer to Comparing LFQ and Isobaric Labeling Strategies in Proteomics Quantification for a more detailed strategy-selection framework.
Intended application |
LFQ LC-MS/MS can support |
A different assay is the better match |
|---|---|---|
Group contrast on independent digests |
Relative change of the same protein |
Amount or copy-number claims |
Open or staggered enrollment |
Later unlabeled runs under one plan |
A closed TMT plex after mixing |
Drug or stimulus response |
Candidate proteins that changed |
Proof of target engagement without orthogonal evidence |
Biofluid or archival tissue |
Unlabeled relative discovery when extraction works |
Metabolic labeling in that matrix |
Short confirmed peptide list |
Not the primary application |
MRM or PRM with a stated calibration model |
How LC-MS/MS Carries Those Applications
LC-MS/MS is the measurement platform behind the applications above, not a second method sitting beside LFQ. Reversed-phase LC spreads peptides in time so that precursor chromatograms can be integrated. MS2 supplies identity, or, in unlabeled DIA, fragment chromatograms that can also carry the quantitative value. Instrument brand, gradient length, and a 300 to 1800 m/z window are laboratory settings. They do not define which biological application is valid.
Load matching, one chromatographic plan, and randomized injection order decide whether an application still has a comparable matrix. Software such as MaxQuant or DIA-NN extracts features. Naming a package does not create a biomarker. Gene ontology or pathway enrichment interprets a differential list. It is not an LC-MS/MS application.
In MtoZ unlabeled discovery offerings, DDA-LFQ is typically reported from MS1 peak-area workflows. Switching the same unlabeled digest to DIA is a separate acquisition decision when completeness, not a new disease area, is the limiting need. Service configuration is not an academic rule that other laboratories must copy.
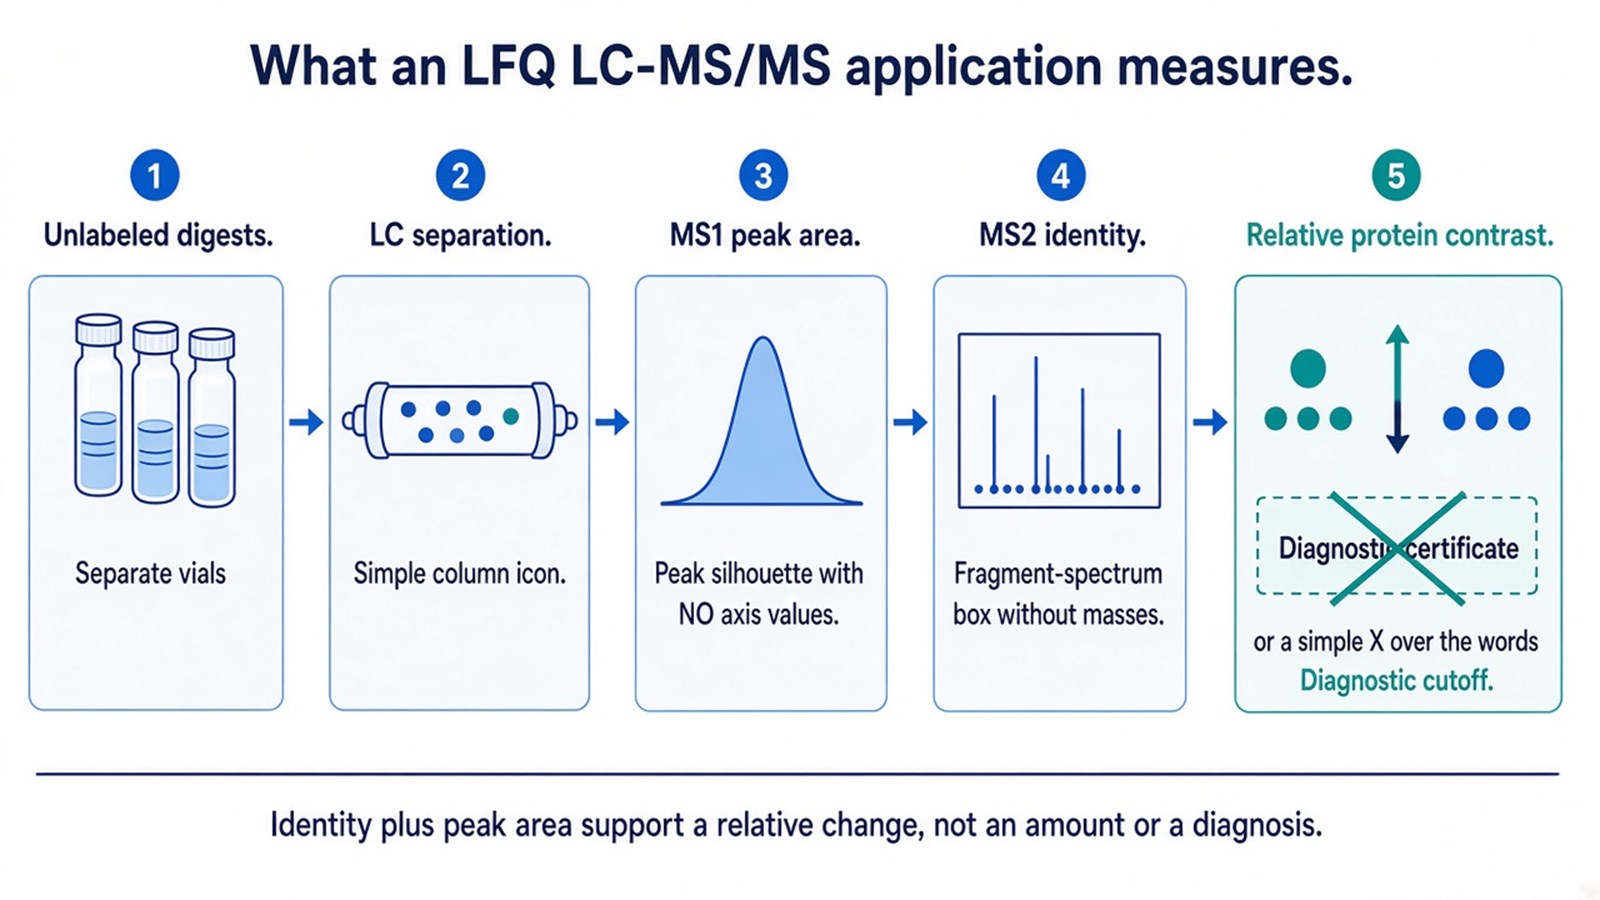
Figure 2. LC-MS/MS applications in LFQ move from unlabeled digests to a relative protein contrast; identity and peak area do not create a diagnostic or absolute result.
What an Application Result Can Support
After alignment, normalization, and protein summarization, an LFQ value can support a relative statement inside the stated contrast. Peptide support, missingness, and run structure belong next to that statement (Cox et al. 2014). A protein that rises after drug exposure can justify a follow-up targeted assay. It cannot, by itself, report mechanism, safety, or a therapeutic dose. A protein that differs between two tissue states can justify a candidate list. It cannot, by itself, support early diagnosis.
LFQ LC-MS/MS applications produce candidate relative changes, not certified biomarkers or environmental verdicts.
Frequently Asked Questions
1. Is biomarker discovery an LFQ application or an LFQ result type?
It is a use of a differential table, not a separate LC-MS/MS method. Candidate proteins from an LFQ contrast still need orthogonal confirmation before anyone treats them as markers.
2. Can LFQ LC-MS/MS be applied to food allergen or pollutant testing as a release assay?
Ordinary discovery LFQ can compare protein profiles when extraction and a database are compatible. It is not a validated allergen or pollutant release test. Those claims need a targeted, calibrated method.
3. Does adding DIA create new LFQ applications?
No. DIA can improve completeness for the same unlabeled contrasts. It does not open diagnostic, agricultural, or multi-omics applications that the relative table could not support under DDA.
4. When is a drug-response study not an LFQ application?
When the needed claim is target occupancy, a calibrated concentration, or a closed multi-group comparison that cannot stay under one chromatographic plan. Those claims point to targeted MS or isobaric labeling.
Once the contrast, unlabeled sample set, and expected protein-level readout are defined, the project can be evaluated as an LFQ LC-MS/MS application. Researchers can review the MtoZ Biolabs Label-Free Quantitative Proteomics Service for project-specific sample evaluation, feasibility assessment, and workflow planning.
