



A high-throughput PPI detection workflow via mass spectrometry is a linked sequence: bait and control design, interaction enrichment, peptide LC-MS/MS, then scored network reporting. The usual capture routes are co-immunoprecipitation–MS (Co-IP-MS), tandem affinity purification–MS (TAP-MS), and crosslinking MS (XL-MS). Label-free or TMT/iTRAQ quantification can sit on the same enrichments when occupancy must be compared across conditions. The workflow reports proteins or crosslinked sites recovered under a stated capture chemistry. It does not, by itself, prove a binary Kd or replace yeast two-hybrid (Y2H) or FRET.
Protein–protein interactions (PPIs) underpin signaling, physiological function, drug-target nomination, and disease-mechanism work. Systems-biology projects need comparable tables across many baits or treatments, which is why mass spectrometry became a central PPI readout. Throughput here means a standardized panel, not an unbounded count of true complexes in one injection. Near-native lysis, quantitative comparison, and optional site mapping are advantages only when controls and processing stay matched across that panel.
Researchers who already have a defined bait panel, treatment contrast, or project objective can also review the MtoZ Biolabs Protein-Protein Interaction Analysis Service for project-specific feasibility and analysis planning.
Why Mass Spectrometry Is Used in a High-Throughput PPI Workflow
Y2H, bimolecular fluorescence complementation (BiFC), and Co-IP remain useful for pairwise screens, cellular reporters, and antibody-based checks. They are limited when the project needs an open, peptide-identified partner list, a condition-to-condition occupancy table, or residue-level contacts. Mass spectrometry changes the readout from one or a few bands to an identified protein or crosslink table and can carry TMT, iTRAQ, SILAC, or label-free comparison on the same chemistry.
Those advantages are design-dependent. A single AP-MS or Co-IP-MS run does not characterize thousands of independent complexes unless the panel, controls, and filters were built for that scale. Native-state language applies to the lysis class, not to every membrane or transient contact. Multi-omics integration can add abundance or modification context. It does not convert an unscored eluate into a validated network.
High-throughput MS PPI work is a repeatable panel workflow. Instrument resolution does not replace matched controls.
Capture Routes Inside the Same Workflow
1. Co-immunoprecipitation–mass spectrometry (Co-IP-MS)
Specific antibodies enrich the target protein and associated complexes; LC-MS/MS then identifies co-precipitated proteins. Co-IP-MS fits stable complexes and targeted questions around a signaling node or core regulator. It depends on antibody specificity, elution control, and a written background plan. Label-free, SILAC, or TMT comparison can be added when the same antibody chemistry is held across conditions.
2. Tandem affinity purification–mass spectrometry (TAP-MS)
Dual tags such as FLAG-HA are fused to the bait, and two sequential affinity steps reduce many non-specific binders (Rigaut et al. 1999). TAP-MS is useful for stable interaction networks and can be run in mammalian, yeast, or insect systems. The second purification also removes weak true partners. Dual-tag TAP is a specificity choice, not a universal upgrade over single-tag AP-MS.
3. Crosslinking mass spectrometry (XL-MS)
Chemical crosslinkers covalently stabilize proximal proteins. Crosslinked peptides are then identified so that interaction sites and conformational constraints can be assigned. Typical reagents include DSSO, BS3, and DMTMM, which differ in spacer length and fragmentation. XL-MS supports complex conformation work and precise site identification. Coverage remains sparse relative to a full interface, and the search is more demanding than a standard Co-IP-MS identification.
4. Quantitative interactome comparison (TMT / iTRAQ)
After Co-IP or TAP, samples can be labeled, pooled, and acquired together so that relative occupancy is compared across drug treatment, cell state, or tissue class. The comparison is valid only when enrichment, digestion, and bait recovery stay matched. TMT or iTRAQ reports relative change of the same bait-associated proteins. It does not report a Kd.
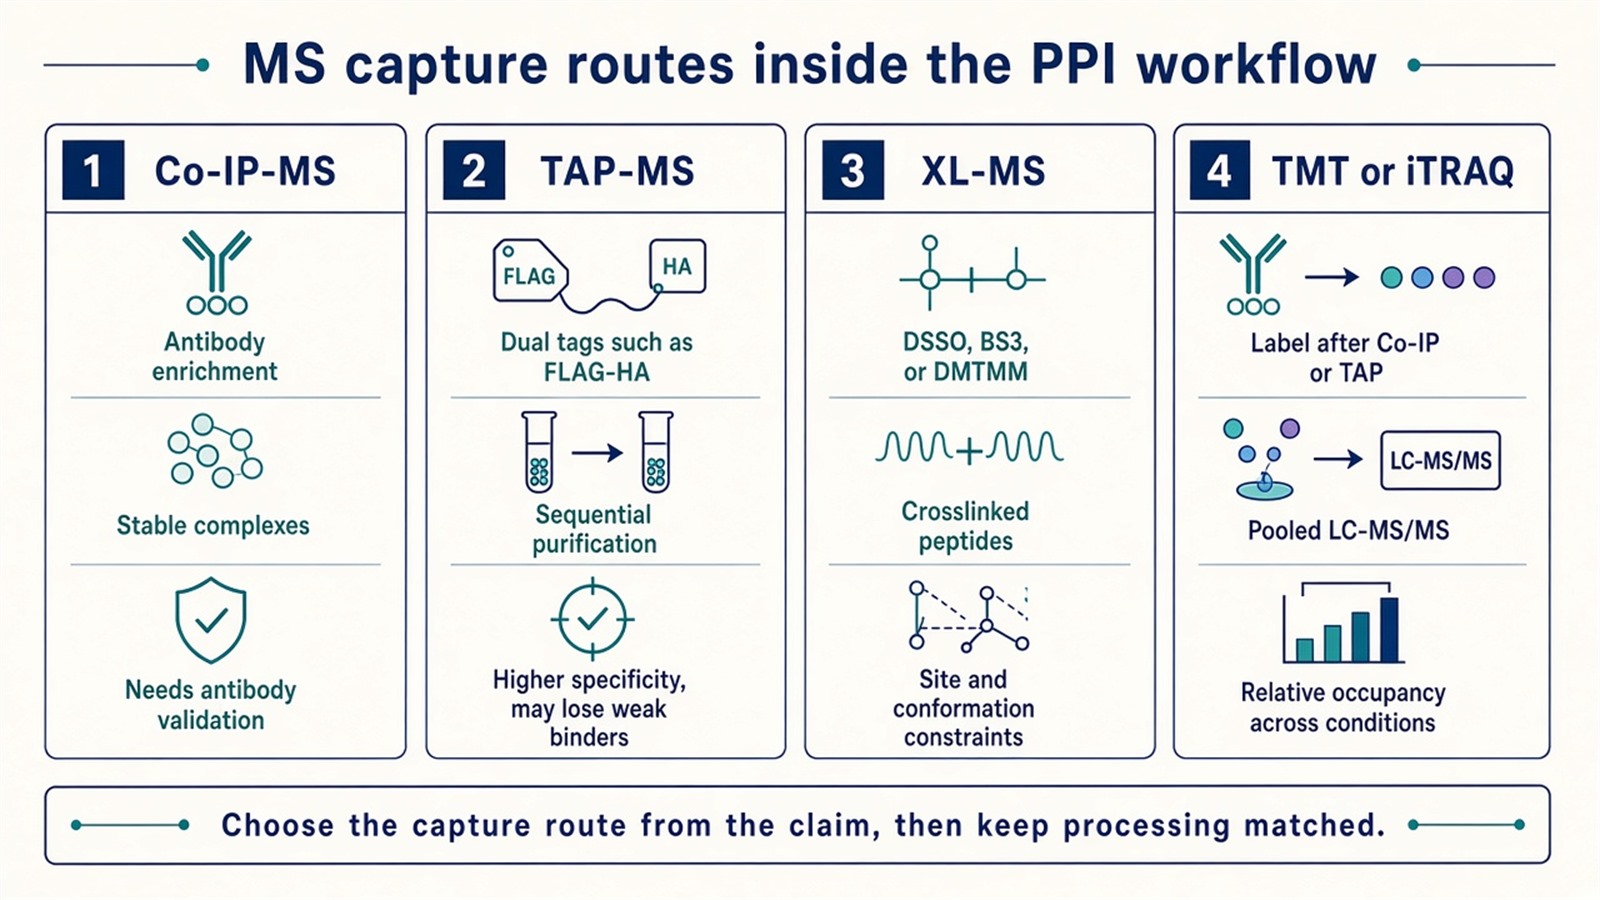
Figure 1. The same LC-MS/MS backend can follow Co-IP-MS, TAP-MS, or XL-MS capture; TMT or iTRAQ compares occupancy only when enrichment stays matched.
The Four-Step High-Throughput Workflow
1. Sample preparation and experimental design
Name the bait, the capture handle (antibody or tag), the control set, and the biological replicates before the first lysis. Decide whether comparison will be label-free or TMT/iTRAQ. Fluorescence reporters belong to a later orthogonal check, not to the labeling plan for a TMT interactome. Controls must share lysis, wash, and digestion with the bait arm.
For which negative-control arms belong in an affinity-MS panel, researchers can refer to AP-MS Controls: Which Negative Controls Do You Need? for more detailed guidance.
2. Interaction enrichment
Run Co-IP, TAP, or chemical crosslinking under a written elution series. Record bead lot, wash stringency, and timing so that a multi-bait panel remains comparable. Background measures that are processed on a different day or with a different resin are not controls for that panel.
3. Protein digestion and mass spectrometry
Convert enriched protein to peptides with a uniform preprocessing route such as FASP or SP3. Acquire high-resolution LC-MS/MS on an Orbitrap-class platform. DDA and DIA are both usable; the choice is a coverage-versus-quantitation trade-off that must be the same across compared arms. On-bead digestion, if used, must be applied to bait and control alike.
4. Bioinformatics and network construction
Identify and quantify proteins with a declared pipeline such as MaxQuant or Spectronaut. Score bait-versus-control enrichment before drawing a network in STRING or Cytoscape. GO or KEGG enrichment and hub ranking can prioritize follow-up. They do not prove that every displayed edge was recovered as a specific interactor in the eluate.
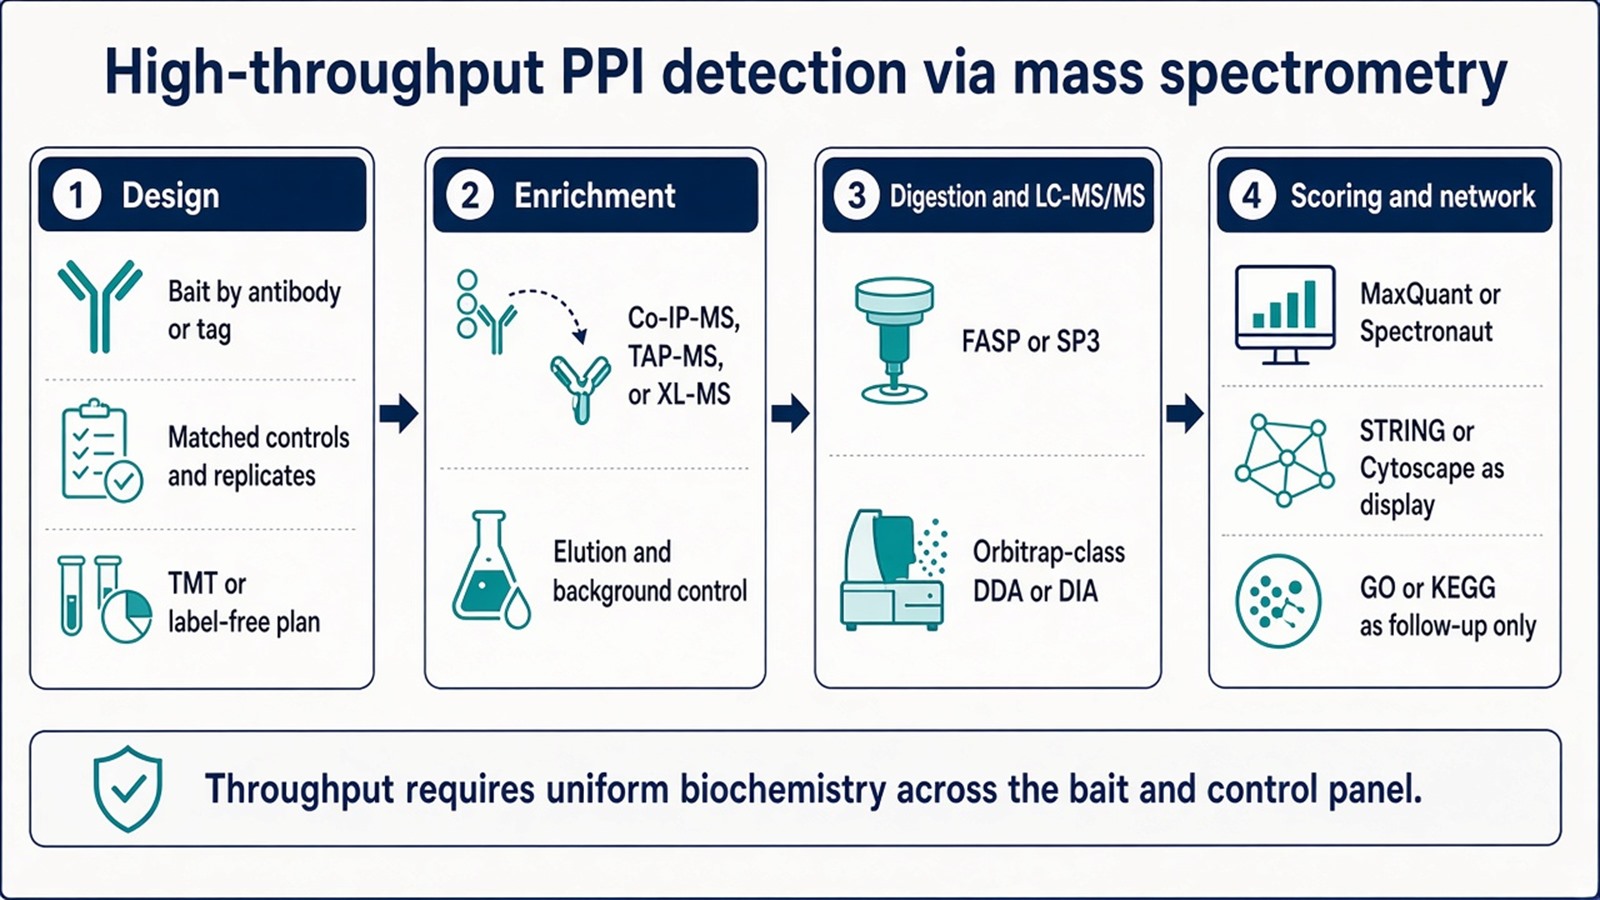
Figure 2. High-throughput PPI detection via mass spectrometry is design, matched enrichment, uniform digestion and LC-MS/MS, then scoring; networks are displays of the scored table.
When the panel is many baits or treatments rather than a single pull-down, High-Throughput Affinity Purification Mass Spectrometry provides a more detailed explanation of batch capture and comparative filtering.
Complementarity with Non-MS Assays
Assay |
What it adds to the MS workflow |
What it does not replace |
|---|---|---|
Y2H |
Pairwise candidate screen |
Cellular co-purification after lysis |
Co-IP / Western |
Selected-edge confirmation |
Open partner discovery |
BiFC or FRET |
Cellular proximity or complementation |
A scored MS occupancy table |
SPR or related chips |
Kd or kinetics on purified pairs |
Lysate complex composition |
A common path is Y2H for a pairwise shortlist, mass spectrometry for cellular complexes or sites, and FRET or a related reporter for selected edges. Complementary methods measure different events. They are not backup copies of the same table.
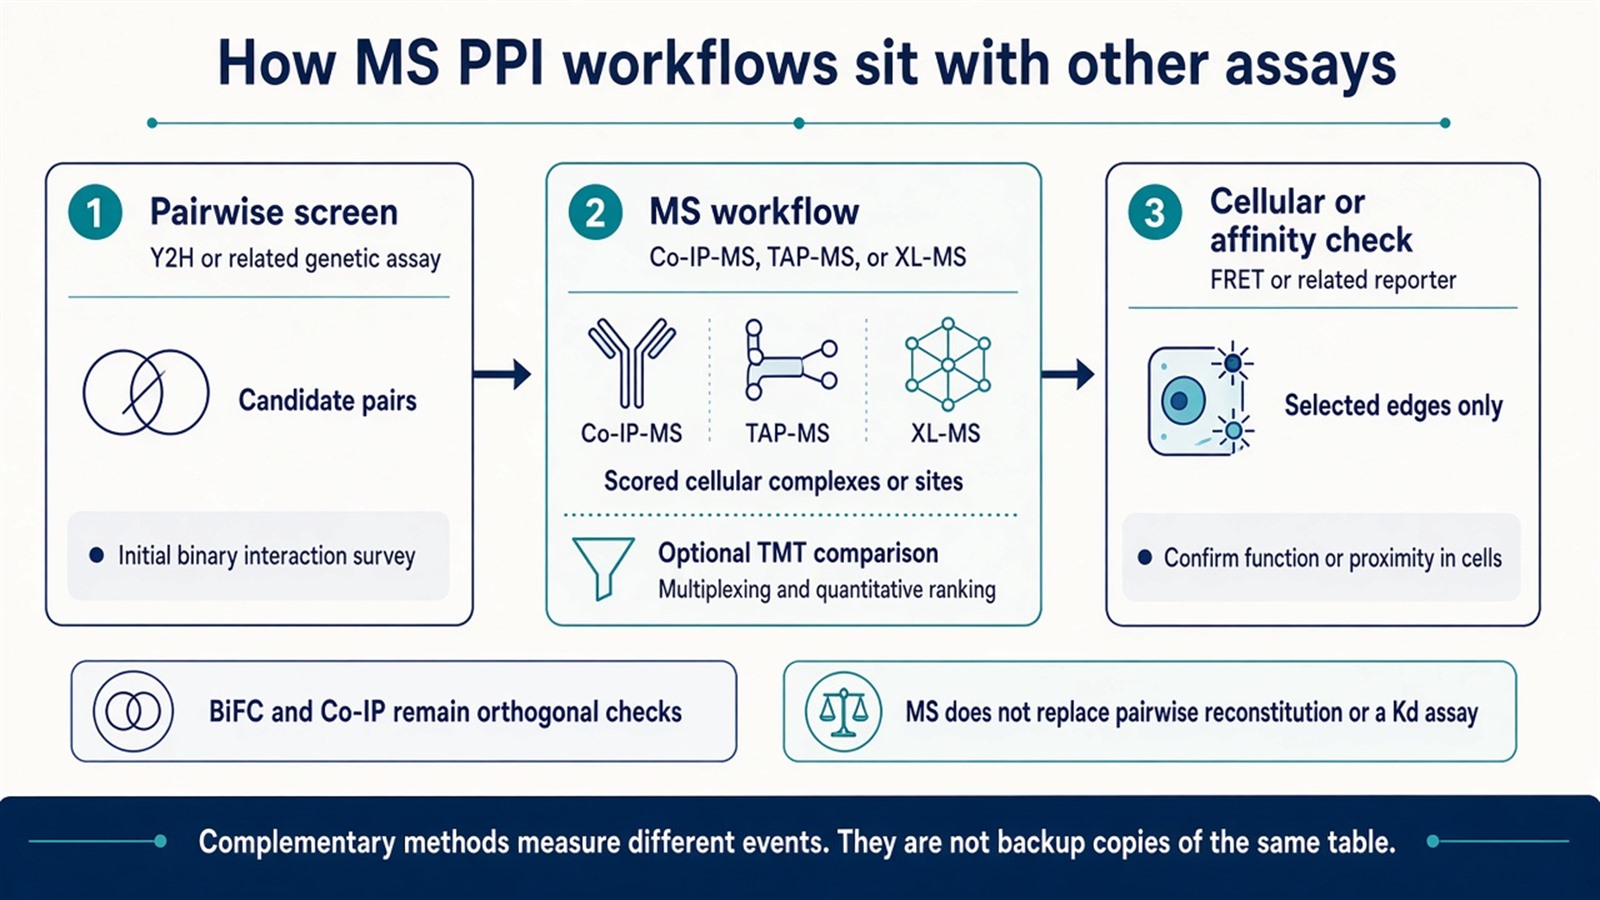
Figure 3. Y2H, MS enrichment, and FRET-class assays are sequenced by evidence type; they do not validate one another as identical interactomes.
For method-level advantages beyond the MS workflow, Advantages and Disadvantages of High-throughput PPI Detection provides a more detailed comparison of Y2H, AP-MS, proximity labeling, and chip assays.
Experimental Limits
Uniform biochemistry across the panel matters as much as instrument capacity. TAP can raise specificity and delete weak true interactors. XL-MS site pairs constrain geometry; they do not reconstruct a complete assembly. TMT compression and missing values still apply. STRING degree is not sample-level proof. Single-cell or AI-assisted mapping can inform later hypotheses. It does not replace the four-step plan on the samples in hand.
Diagnostic or companion-diagnostic claims sit outside ordinary research PPI-MS reporting unless a separately validated assay framework is in place.
Project Planning and Service Scope
A usable high-throughput PPI-MS plan names the capture route, control set, replicate structure, digestion chemistry, acquisition mode, and quantification method. MtoZ Biolabs can scope labeling-strategy design, enrichment, Orbitrap-class acquisition, and scored interpretation for Co-IP-MS, TAP-MS, XL-MS, and TMT comparison. Oncology, neuroscience, or immunology context does not change those design requirements.
Once the capture route and panel rules are defined, the project can be scoped as a high-throughput PPI mass spectrometry study. Researchers can review the MtoZ Biolabs Protein-Protein Interaction Analysis Service for project-specific sample evaluation, feasibility assessment, and workflow planning.
Frequently Asked Questions
1. Is Co-IP-MS the same as AP-MS?
They share LC-MS/MS identification of co-enriched proteins. Co-IP-MS usually starts from an antibody against the endogenous or tagged bait. AP-MS is the broader tag- or antibody-based family. TAP-MS is a dual-tag subset.
2. Does high-throughput mean thousands of true complexes in one run?
No. Throughput refers to a standardized multi-bait or multi-condition panel. The number of defensible interactors depends on controls, bait recovery, and scoring.
3. When should TAP-MS be chosen over single-step Co-IP-MS?
Choose TAP-MS when a second purification is needed to cut background on a stable complex. Do not choose it if weak partners are the main target.
4. Can TMT be added to every Co-IP-MS or TAP-MS workflow?
Yes, when enrichment and bait recovery are matched and the claim is relative occupancy. TMT does not convert the table into affinity constants.
5. Do DDA and DIA change the biological claim?
They change coverage and missing-value behavior. They do not change the requirement for matched controls and a declared score.
Reference
- Gingras A.C., Gstaiger M., Raught B., Aebersold R. (2007). Analysis of protein complexes using mass spectrometry. Nat. Rev. Mol. Cell Biol., 8, 645-654. https://doi.org/10.1038/nrm2208
- Rigaut G., Shevchenko A., Rutz B., et al. (1999). A generic protein purification method for protein complex characterization and proteome exploration. Nat. Biotechnol., 17, 1030-1032. https://doi.org/10.1038/13732
- Leitner A., Walzthoeni T., Kahraman A., et al. (2010). Probing native protein structures by chemical cross-linking, mass spectrometry, and bioinformatics. Mol. Cell. Proteomics, 9, 1634-1649. https://doi.org/10.1074/mcp.R000089-MCP201
- Thompson A., Schafer J., Kuhn K., et al. (2003). Tandem mass tags: a novel quantification strategy for comparative analysis of complex protein mixtures by MS/MS. Anal. Chem., 75, 1895-1904. https://doi.org/10.1021/ac0262560
