LC-MS Analysis of Pull-down Proteins
MtoZ Biolabs helps researchers identify proteins recovered in pull-down experiments and investigate potential binding partners of a target protein.
From tagged bait protein capture assessment to Orbitrap LC-MS/MS analysis, our service supports protein identification and comparison of pull-down results across experimental conditions.
- Tagged Bait Capture Strategy Assessment
- Orbitrap-Based LC-MS/MS with Mass Accuracy Below 5 ppm
- Protein Identification and Comparative Abundance Analysis
What Is LC-MS Analysis of Pull-down Proteins?
Pull-down MS combines affinity-based protein capture with liquid chromatography–tandem mass spectrometry (LC-MS/MS). It is used to identify proteins recovered with a bait protein and generate candidate binding partners for subsequent investigation.
The approach is particularly useful when a study involves multiple potential binding partners and requires protein identification beyond a predefined list of candidates. The quality of the resulting candidate list depends on the bait protein, capture conditions, background binding, and experimental controls.
When LC-MS Analysis of Pull-down Proteins Is a Good Fit
Pull-Down-MS is a suitable option when a study requires protein identification following affinity-based capture, particularly in the following situations:
1. Screening Unknown Binding Partners
Identify proteins recovered with a defined bait protein when candidate interactors have not been established.
2. Investigating Fusion-Tagged Proteins
Evaluate suitable GST-, His-, MBP-, or Strep-tagged bait constructs using a compatible affinity capture strategy.
3. Analyzing Completed Pull-Down Experiments
Identify proteins in enriched magnetic beads, eluates, gel bands, or compatible protein solutions.
4. Comparing Experimental Conditions
Examine differences in protein intensity between bait-containing samples and controls when the study design supports relative abundance comparison.
5. Prioritizing Candidate Interactors
Use protein identification results to select candidates for subsequent validation or functional investigation.
The bait protein is an important consideration. Pull-Down-MS works best when the bait can be obtained in a form that retains the binding activity relevant to the experiment. If binding depends on a post-translational modification absent from the recombinant protein, or if a membrane protein cannot be maintained in a suitable soluble form, another experimental strategy may be needed.
For projects focused on protein associations under cellular or native conditions, Co-IP may be more appropriate. If the main objective is to identify a target protein and characterize associated modifications, IP-MS may be a better fit.
Pull-Down MS vs Co-IP vs IP-MS: How to Choose?
The appropriate method depends on the biological system, the starting material, and the evidence required.
|
Decision factor |
Pull-Down MS |
||
|
Typical research system |
Suitable in vitro binding systems |
Cellular or native protein complexes |
Immunoaffinity enrichment of a target protein and associated material |
|
Starting material |
Purified bait protein and compatible binding partners, or completed pull-down material |
Suitable cellular or biological material and an appropriate antibody |
Suitable target-containing material and an appropriate immunoaffinity strategy |
|
Main purpose |
Identify proteins captured by a bait protein |
Investigate protein associations in a biological context |
Identify proteins associated with an immunoprecipitated target |
|
Important consideration |
Bait activity, capture conditions, and background binding |
Antibody specificity, target abundance, and preservation of protein complexes |
Enrichment specificity, target recovery, and background proteins |
Which service should you choose?
- Choose Pull-Down MS when the study uses a suitable bait protein for an in vitro capture experiment and requires identification of recovered proteins.
- Consider Co-IP when the primary question concerns protein association under cellular or native conditions.
- Consider IP-MS when the objective is to identify a target protein and its associated proteins or modifications through immunoaffinity enrichment.
These methods provide different types of evidence and are not interchangeable. If the research objective or starting material is unclear, provide the project details for assessment before selecting a workflow.
Service at MtoZ Biolabs
MtoZ Biolabs provides downstream LC-MS analysis for pull-down-derived samples. The service covers sample preparation, mass spectrometry acquisition, database searching, and result reporting.
The service can accommodate the following sample formats, subject to sample suitability:
1. Gel Bands
Protein bands visible to the naked eye.
2. Pull-Down or IP Magnetic Beads
Enriched bead-bound material, with sample quantity and buffer composition confirmed before submission.
3. Eluates
Protein material recovered from affinity-based capture.
4. Protein Solutions
Compatible protein solutions, including samples processed in loading buffer.
5. Original Biological Samples
Animal or plant tissues, cells, microorganisms, and body fluids may be considered after project assessment. The required capture or sample-processing strategy depends on the research objective, and an appropriate protein reference database must be available.
For projects requiring both bait-protein capture and mass spectrometry, the experimental scope should be confirmed before submission. A completed pull-down sample and an original biological sample are not interchangeable starting materials.
Analysis Workflow
1. Project and Sample Assessment
Confirm the research objective, bait-protein identity, affinity tag, sample format, protein amount, buffer composition, and reference database.
2. Sample Preparation
Prepare the submitted material for proteomic analysis according to its format.
3. Protein Digestion
Perform reduction, alkylation, and tryptic digestion to generate peptides.
4. C18 Desalting
Remove interfering components before LC-MS/MS analysis.
5. Nano-LC-MS/MS Analysis
Separate peptides by nano-liquid chromatography and acquire high-resolution mass spectra using an Orbitrap-series platform.
6. Database Searching
Process the acquired data using MaxQuant and an appropriate UniProt or customer-provided reference database.
7. Data Reporting
Compile protein and peptide identifications, sequence coverage, identification scores, intensity information, and the PDF report.
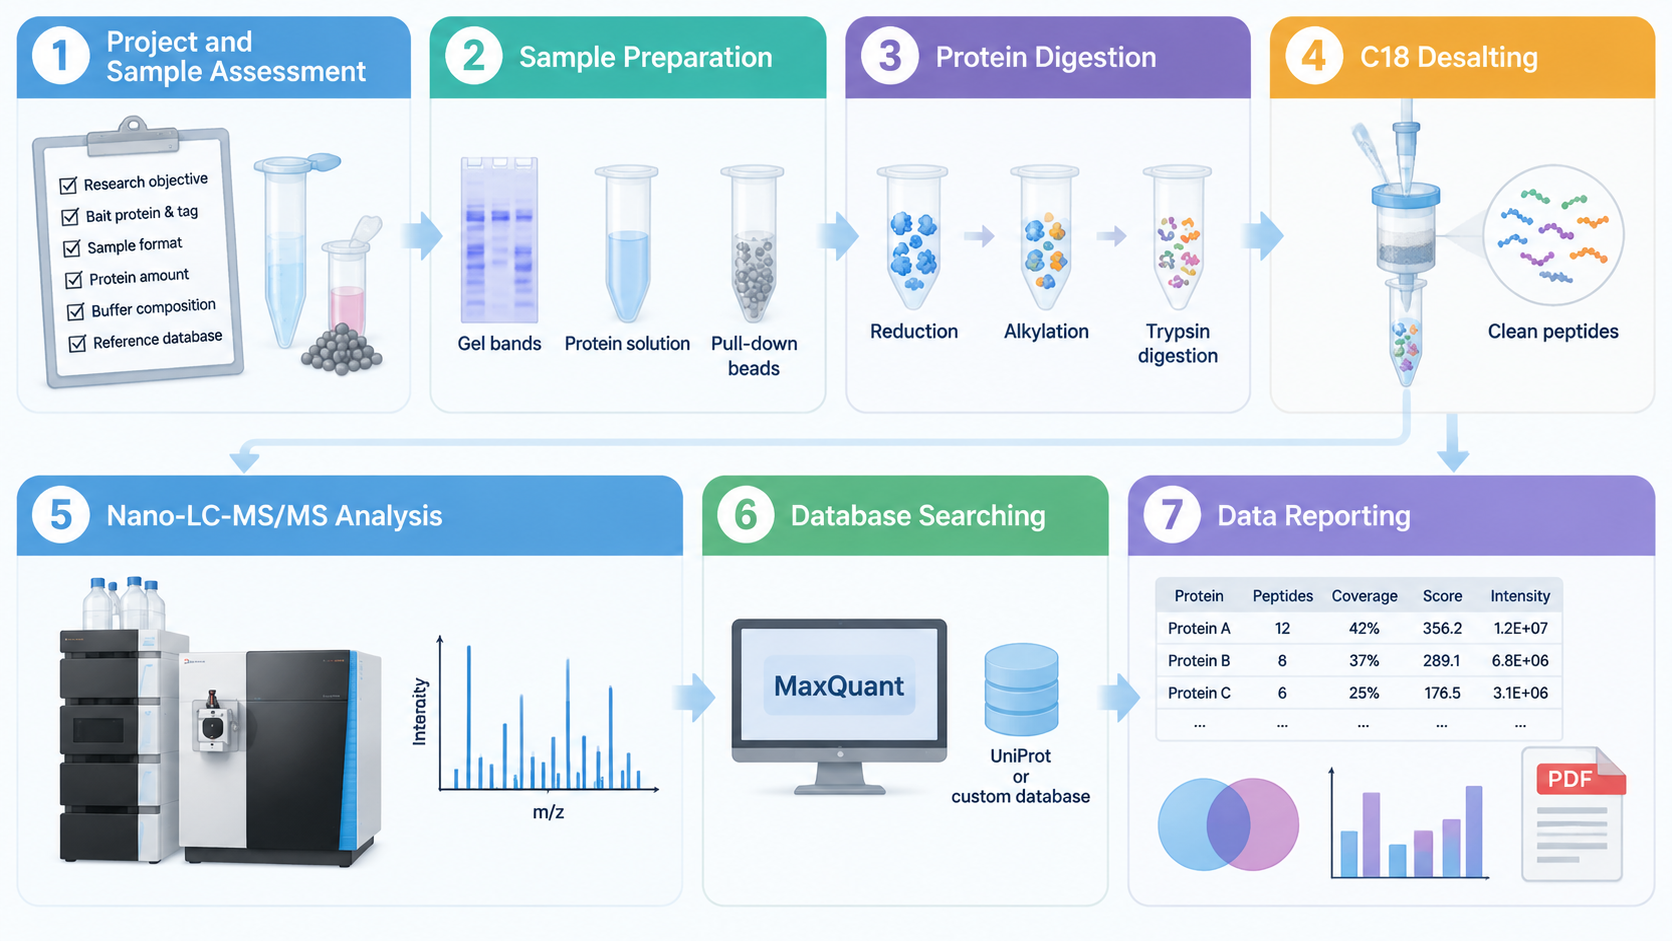
Figure1. LC-MS Analysis Workflow of Pull-down Proteins
Experimental Instruments
1. Mass Spectrometry Platform
Orbitrap-series high-resolution mass spectrometry.
2. Mass Accuracy
Below 5 ppm.
3. Primary Acquisition Mode
Data-dependent acquisition (DDA).
4. Alternative Acquisition Mode
Data-independent acquisition (DIA) may be considered upon request and project assessment.
5. Data Analysis Software
MaxQuant.
6. Reference Database
UniProt or a suitable customer-provided custom database.
Sample Submission Requirements
|
Sample type |
Submission requirements |
Storage and transport |
|
Gel bands |
Protein bands must be visible to the naked eye |
Ship at room temperature or with an ice pack |
|
Protein solutions or eluates |
Total protein ≥20 μg; concentration >0.5 μg/μL |
Rapidly freeze in liquid nitrogen and ship on dry ice |
|
Pull-down or IP magnetic beads |
Confirm sample amount and buffer composition before submission |
Rapidly freeze in liquid nitrogen and ship on dry ice |
|
Original biological samples |
Confirm sample suitability, project scope, and reference database availability |
Confirm preparation and transport requirements before shipment |
1. SDS
The concentration must be below 0.1%.
2. Salt and Detergents
High-salt samples and samples containing high concentrations of detergents must be cleaned up before submission.
3. Sample Integrity
Severely degraded samples are not accepted.
4. Reference Database
Provide the organism or sample source and relevant protein sequence information. Customer-provided custom databases are accepted when suitable.
5. Non-Standard Samples
Confirm the sample format, buffer composition, and project scope before shipment.
For original tissues, cells, microorganisms, or body fluids, the sample amount and preparation requirements depend on the proposed experimental route. Contact the project team before shipment to determine whether additional capture or sample-processing steps are needed.
Advantages of Pull-Down MS
1. Candidate Interactor Screening
Identify proteins recovered with a suitable fusion-tagged bait, including candidates that were not specified in advance.
2. High-Resolution Protein Identification
Orbitrap-based LC-MS/MS provides peptide fragmentation data for database searching and protein identification.
3. Peptide-Level Evidence
Protein assignments are accompanied by peptide identifications, sequence coverage, and identification scores.
4. Flexible Analytical Strategy
Evaluate suitable bait-protein capture projects and analyze completed pull-down-derived samples.
5. Relative Abundance Comparison
Compare intensity data across suitable experimental groups when supported by the study design.
6. One-Time Charge
Our pricing is transparent, with no hidden fees or additional costs.
Deliverables
1. Raw Mass Spectrometry Files
Original data generated during LC-MS/MS analysis.
2. Protein and Peptide Identification Excel Files
Protein and peptide assignments, sequence coverage, identification scores, and intensity information.
3. Relative Abundance Data
Intensity-based comparison results when supported by the experimental design.
4. PDF Report
A summary of the analysis and identification results.
FAQ
Q1. Can MtoZ Biolabs assess a pull-down strategy if I have not completed the experiment?
For suitable customer-provided bait proteins, a tag-antibody-based capture strategy can be evaluated. Please provide the bait-protein or construct information, affinity tag, preparation status, and research objective so the appropriate workflow can be assessed.
Q2.Can Pull-Down-MS prove direct binding between the bait and a candidate protein?
No. The analysis identifies proteins recovered under the tested conditions. Direct binding requires additional evidence, and nonspecific enrichment or indirect association should be considered when interpreting candidate proteins.
Q3.Can Pull-Down-MS confirm that an interaction occurs inside cells?
An in vitro pull-down result alone cannot establish that an interaction occurs under physiological cellular conditions. For studies focused on cellular protein associations, Co-IP or another appropriate validation strategy may be considered.
Q4. Can original tissues or cells be submitted instead of completed pull-down samples?
Selected original biological samples may be considered, including animal or plant tissues, cells, microorganisms, and body fluids. The appropriate workflow depends on the research objective, sample properties, and availability of a suitable protein reference database. Project suitability should be confirmed before shipment.
Start Your Project with MtoZ Biolabs
To discuss your Pull-Down-MS project, provide the bait-protein identity and affinity tag, construct or preparation information, sample type, buffer composition, available controls, and research objective.
If you already have a completed pull-down sample, include the sample format and estimated protein amount. If the experiment is still being planned, describe the intended binding question and whether a suitable bait protein is available. This information helps our team assess the appropriate capture and LC-MS/MS workflow for your study.
MtoZ Biolabs, an integrated chromatography and mass spectrometry (MS) services provider.
Related Services
SILAC Based Co-IP-MS for Protein Interaction Analysis Service
Demo for LC-MS Analysis of Pull-down Proteins
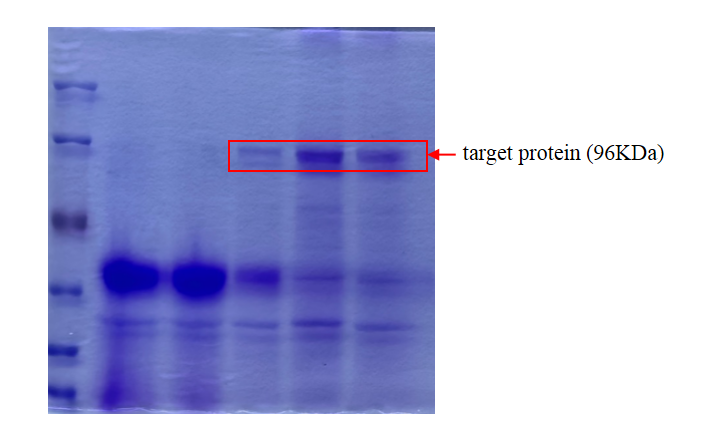
SDS-PAGE Analysis of Enriched Protein Samples
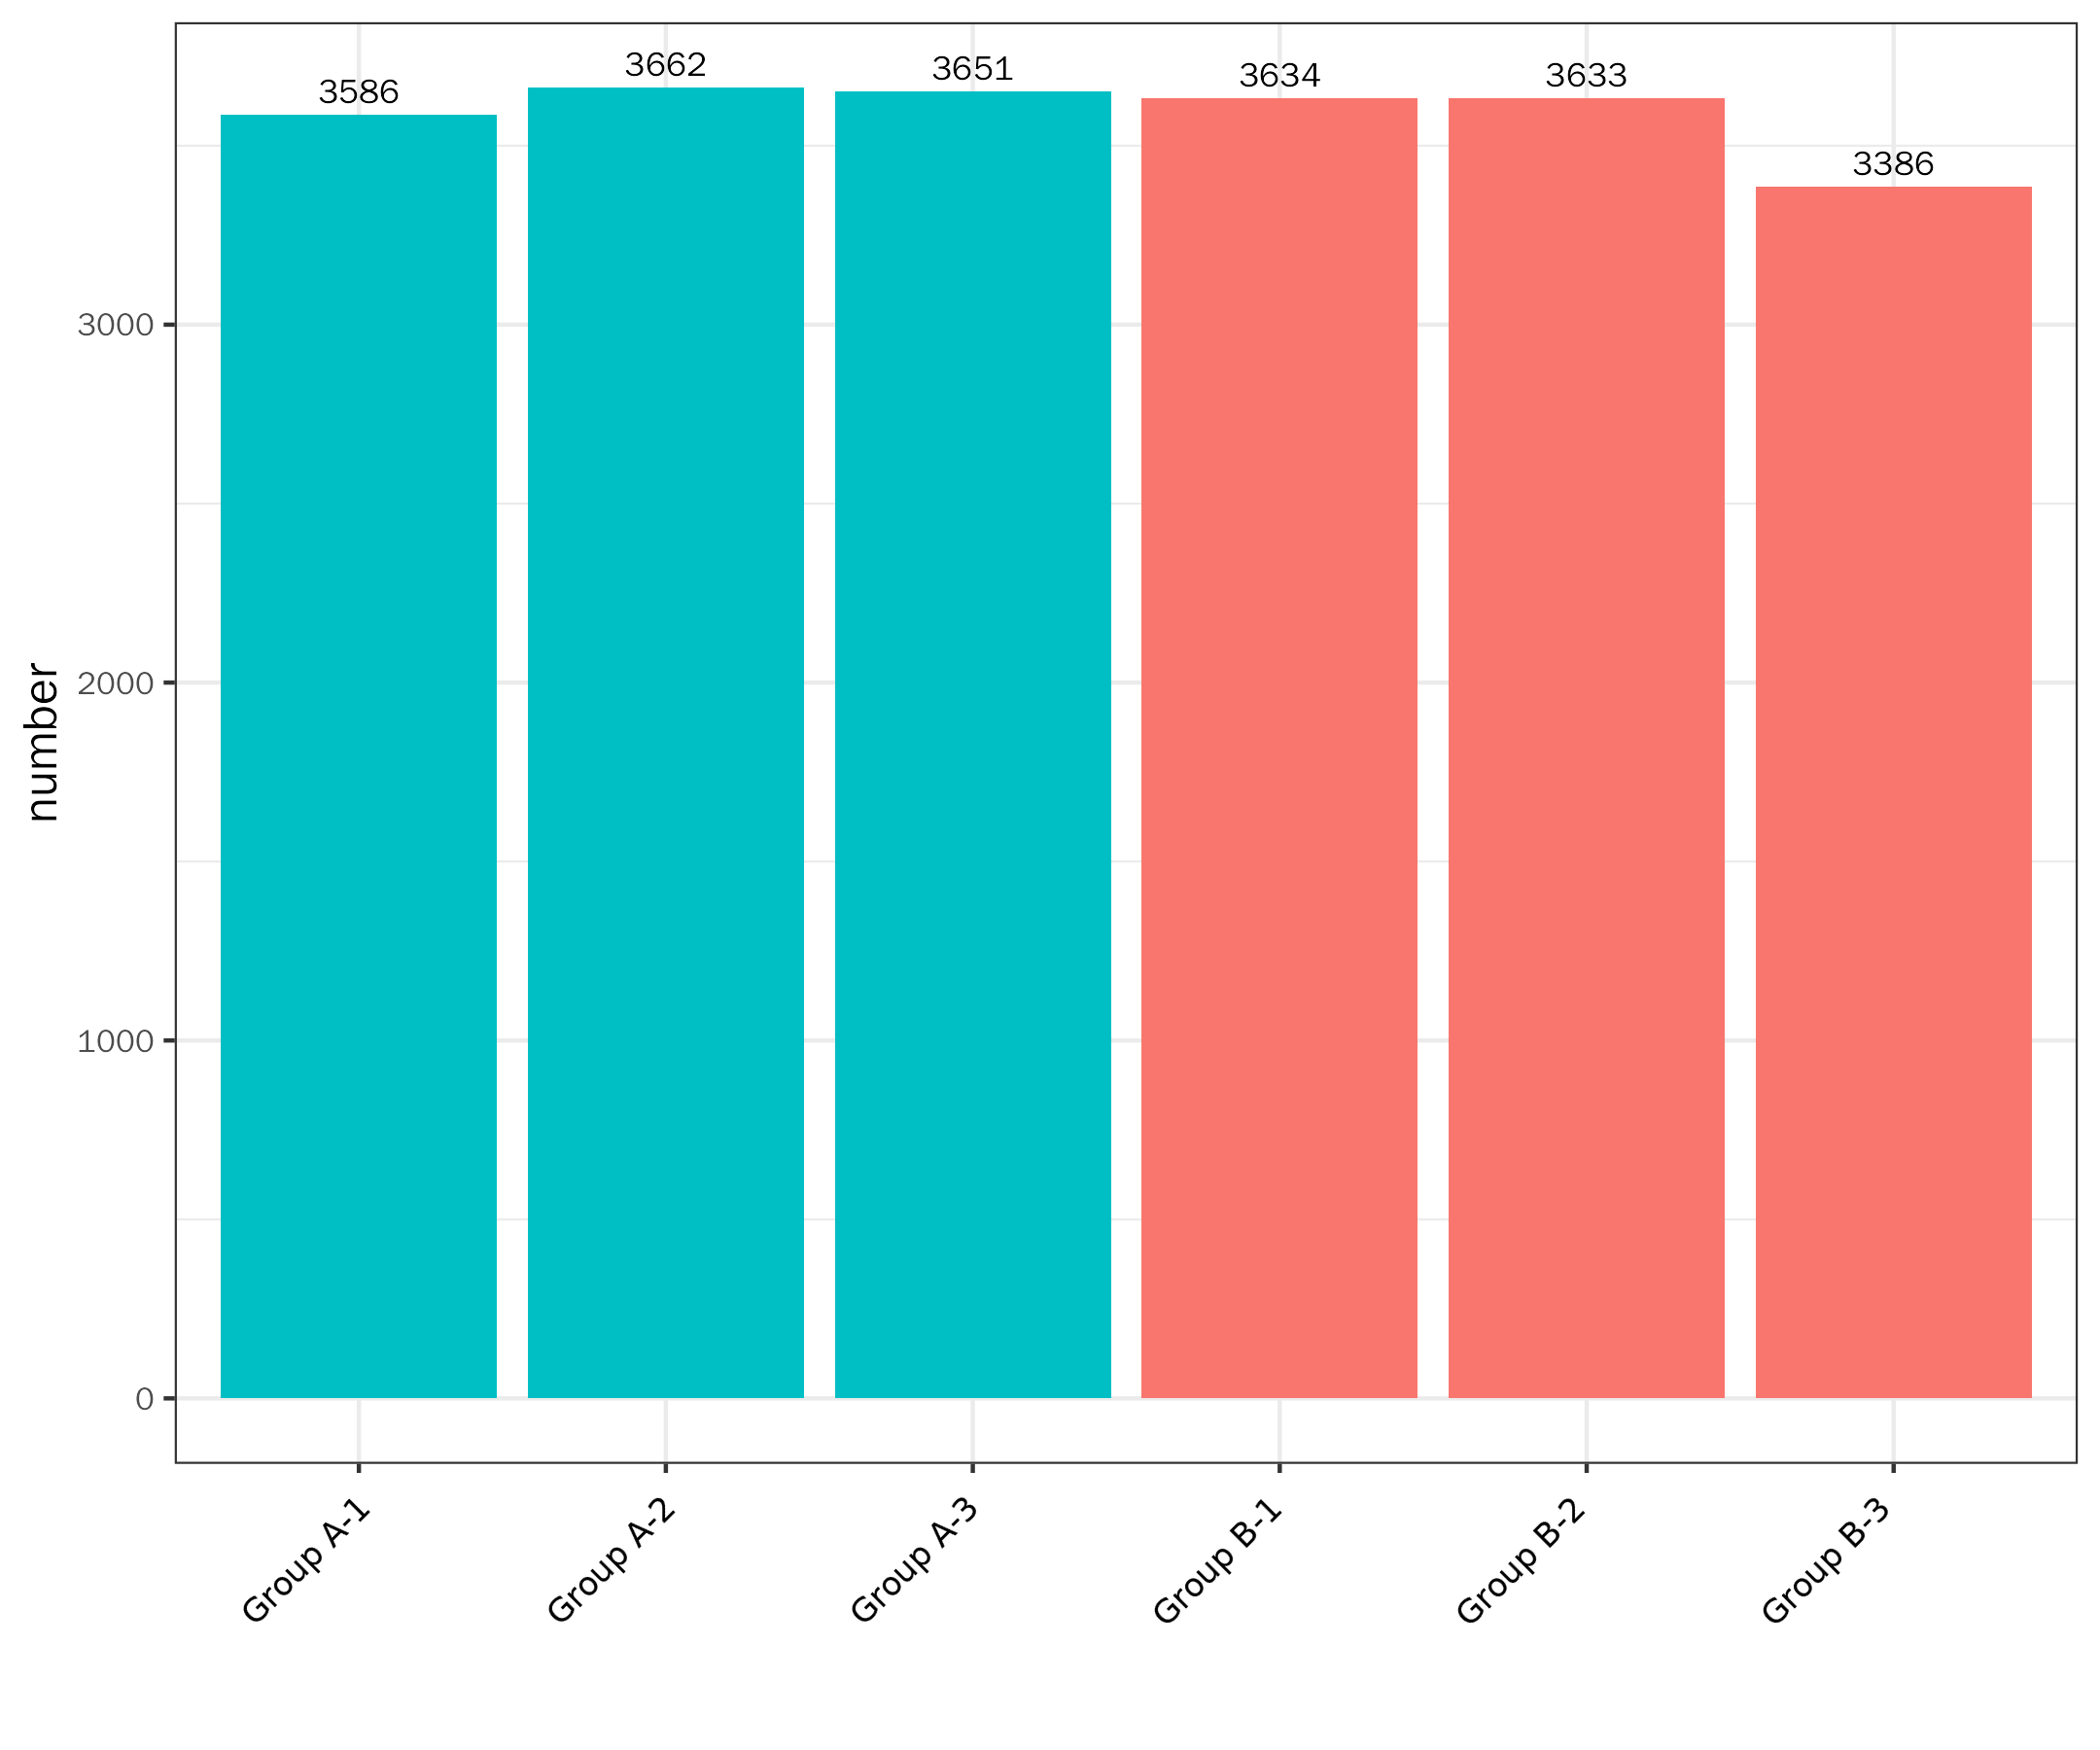
Protein Identification Counts Across Samples
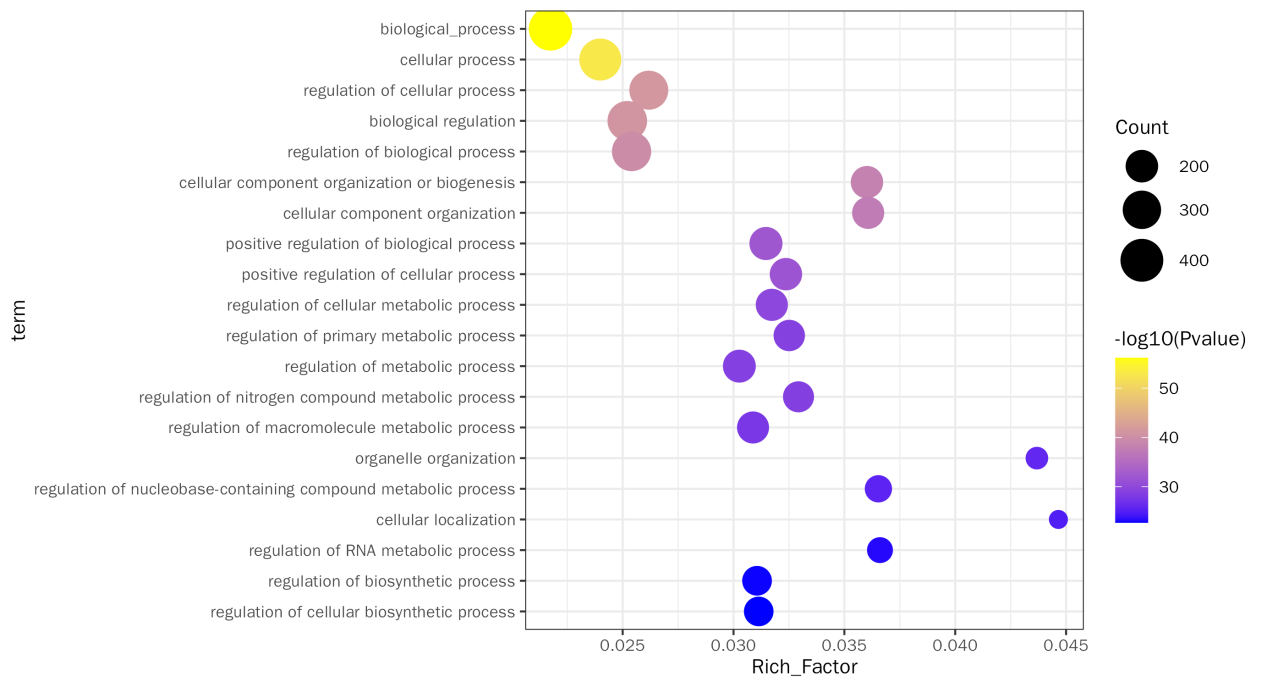
GO Enrichment Bubble Plot
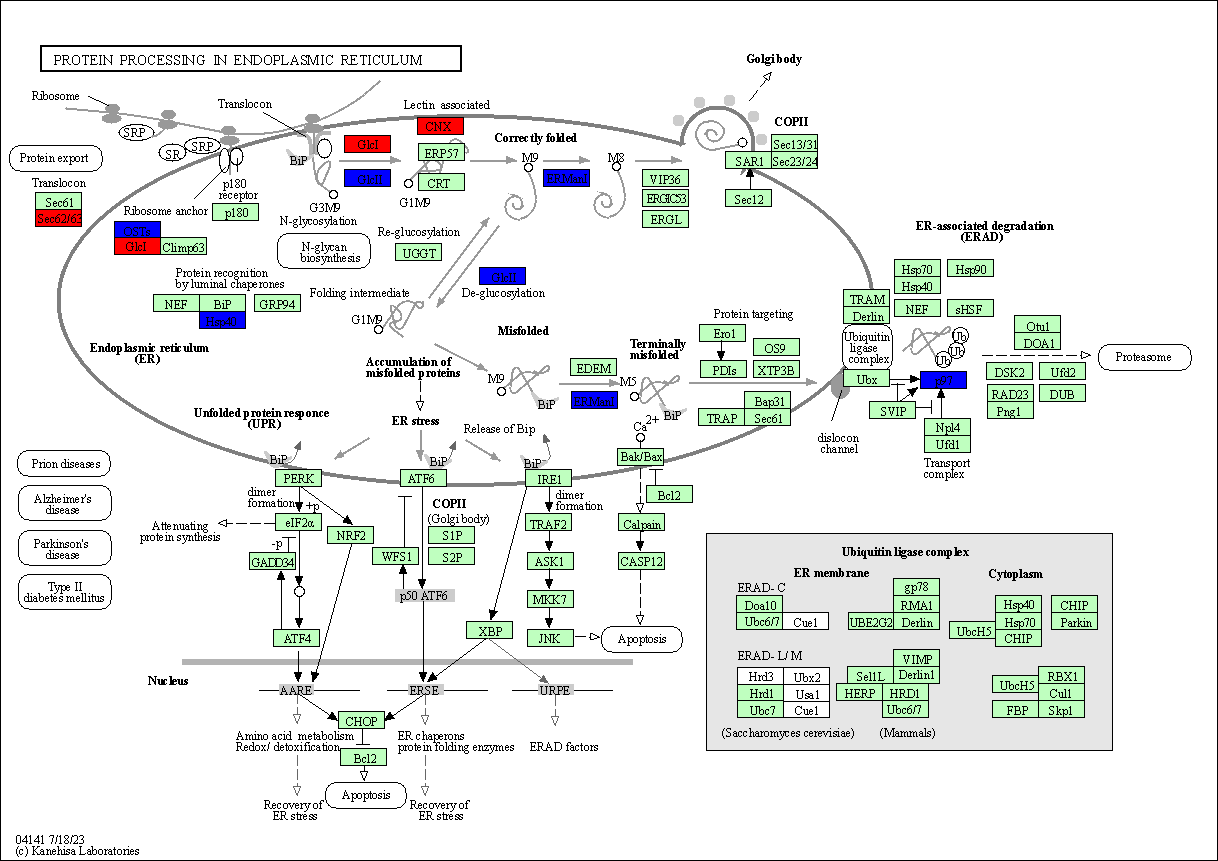
KEGG Pathway Diagram
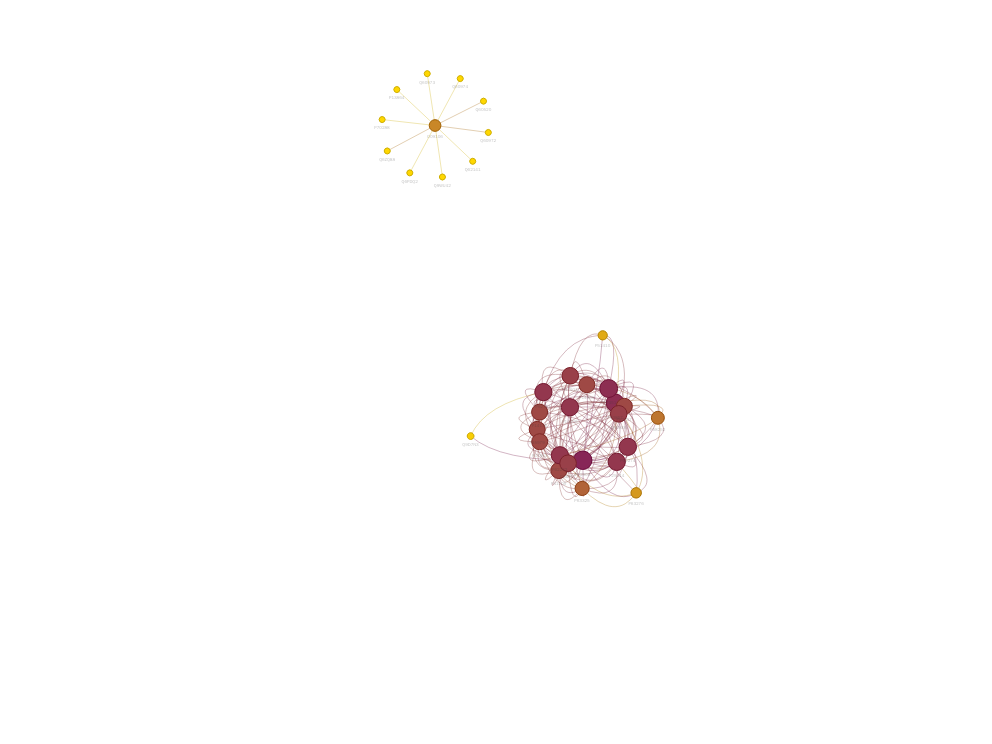
PPI Network Highlighting Hub Proteins