Overview of Targeted Proteomics: Methods, Techniques, Key Points, and Applications
Targeted proteomics means quantifying a closed set of proteins through predefined proteotypic peptides on liquid chromatography-tandem mass spectrometry (LC-MS/MS). Multiple reaction monitoring (MRM) or selected reaction monitoring (SRM) records scheduled precursor-to-product transitions on a triple quadrupole (Lange et al. 2008). Parallel reaction monitoring (PRM) isolates scheduled precursors on a high-resolution instrument and stores product-ion spectra for fragment review (Peterson et al. 2012). The defining rule is that targets are chosen before acquisition. The method does not invent an open proteome map from the same file.
Relative peak ratios answer fold-change under a declared normalization rule. Absolute amount needs stable isotope-labeled internal standard (SIS) peptides or Absolute QUAntification (AQUA) design plus a calibration model in the study matrix (Gerber et al. 2003; Carr et al. 2014). Surrogate peptides report the measured sequence under the stated digest. They do not represent every isoform or modified form by default (Bantscheff et al. 2012). Data-independent acquisition (DIA) or SWATH-style windows can support retrospective mining, but they belong to discovery-class acquisition when the analyte list is not closed before the run.
In MtoZ Biolabs targeted offerings, method choice is scoped after list status, matrix notes, and claim unit are written in the brief. Service intake appears in Targeted Proteomics Service.
What Targeted Proteomics Means in Project Language
Four definition layers belong in the brief and stay separated.
List layer states that proteins or peptides are named and versioned before acquisition. An open candidate pool is still discovery work.
Acquisition layer chooses MRM/SRM for stable scheduled transitions or PRM when high-resolution fragment confirmation is required.
Quantitation layer separates relative normalization from SIS-calibrated absolute units.
Claim layer matches interpretation to the declared purpose: cohort remeasurement, pathway contrast, or concentration reporting. Clinical diagnosis or release language needs a separately validated method beyond a research panel.
Targeted proteomics remeasures a closed peptide panel. Prior knowledge of the targets is a requirement, not a weakness to hide.
How MRM, PRM, and Discovery-Class Methods Differ
Method names should not be treated as synonyms.
MRM/SRM spends dwell time on predefined transitions and fits high-throughput remeasurement when matrix-matched ion ratios stay stable (Lange et al. 2008). PRM records a full high-resolution fragment set for each scheduled precursor, which helps when unit-resolution transitions remain interference-limited (Peterson et al. 2012; Gallien et al. 2012). Hybrid panels keep stable peptides on MRM and hard peptides on PRM.
Discovery data-dependent acquisition (DDA) or DIA profiles an open or semi-open proteome before any panel is frozen. Those routes nominate candidates. They do not replace a locked targeted method for the same decision. Treating DIA as core targeted acquisition confuses list status with post-acquisition filtering. Route timing between open profiling and closed-panel remeasurement appears in Targeted Mass Spectrometry or Discovery LC-MS/MS? Matching MRM, PRM, and Profiling Routes to Your Analytical Goal. Platform detail appears in Parallel Reaction Monitoring vs MRM: Which Fits?.
Key operating points are shared across targeted methods: proteotypic uniqueness, matrix-matched assay build, matched preparation across contrast arms, and a declared relative or absolute claim. Absolute reporting intake appears in Absolute Quantitative Analysis (AQUA) Service.
End-to-End Workflow From Definition to Operable Assay
A practical definition-to-assay spine usually runs as follows.
- Freeze the analytical question: remeasure named proteins, confirm a pathway panel, or report absolute amount.
- Confirm list status: closed peptide panel versus still-open proteome.
- Select proteotypic surrogates that are unique, detectable, and stable under the planned digest.
- Declare relative normalization or SIS/AQUA absolute rules and spike timing.
- Choose MRM/SRM, PRM, or a hybrid design from matrix risk and fragment-review need.
- Match extraction, digest, cleanup, and load across every contrast arm.
- Build transitions or fragment rules on matrix-matched digests; run a pilot before full cohort shipment.
- Acquire interleaved study samples with blanks, quality control (QC) pools, and calibrators when absolute units are required.
- Integrate peaks, apply inclusion rules, flag failed targets, and archive panel version with the report.
Steps 1 through 4 protect the definition and claim. Steps 5 through 9 are the method, prep, and reporting record. MRM/SRM service context appears in SRM-MRM Quantitative Proteomics Services. PRM service context appears in Parallel Reaction Monitoring (PRM) Service. Operational biomarker and pathway detail appears in Targeted Proteomics Workflows for Biomarker and Pathway Quantitation.
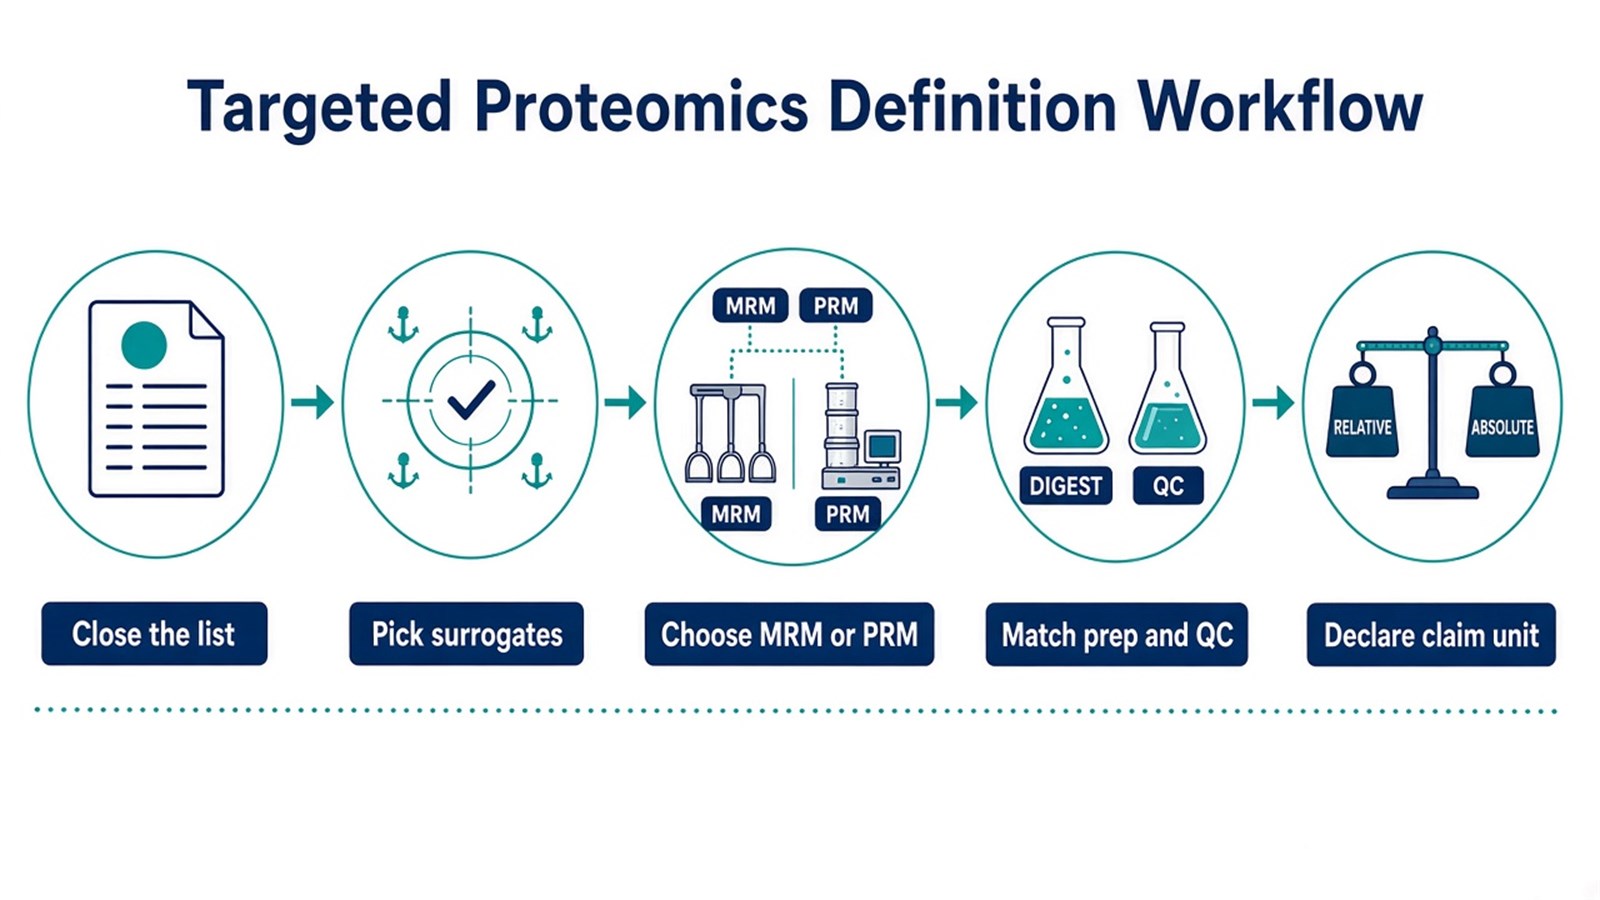
Figure 1. Targeted proteomics is defined by a closed list and selective acquisition; claim unit and matched prep must be locked before cohort measurement.
Key Points and Parameters That Must Be Declared
Layer |
Parameter that must be declared |
Failure mode if ignored |
|---|---|---|
Definition |
Closed panel versus open proteome |
Discovery question run as targeted |
Surrogate |
Proteotypic uniqueness and digest rules |
Protein-level overclaim from one peptide |
Method |
MRM/SRM, PRM, or hybrid rationale |
Wrong specificity for matrix risk |
Claim unit |
Relative versus SIS absolute |
Concentration language without calibrators |
Matrix |
Type, storage, pretreatment parity |
Suppression read as biology |
Cycle time |
Retention windows and dwell schedule |
Dropped targets on dense panels |
QC plan |
Blanks, pools, precision checks |
Batch drift unread |
SIS or AQUA |
Spike step and assigned amount |
Absolute claim without standard design |
Inclusion rules |
Failed-target handling |
Silent zeros in group means |
Deliverable |
Table, method notes, QC flags |
Numeric file without assay context |
Limits follow the same table. Targeted methods cannot quantify unnamed proteins without peptide selection. Assay development is upfront work, not optional overhead. Peptide detectability varies by matrix. A larger panel does not replace a matrix pilot. Clinical translation language needs separate validation design beyond research remeasurement.
What Targeted Proteomics Can Support
A locked relative panel supports same-peptide comparison across samples acquired with one method version. A locked absolute panel supports concentration or amount for surrogates inside the calibrated interval when SIS design and QC recovery are documented.
Biomarker candidate remeasurement and pathway panel tracking are common fit cases after the list is closed. Failed targets support exclusion or redesign. They do not support silent zeros. Cross-study comparison needs matching normalization or calibration assumptions.
Targeted proteomics supports remeasurement of known targets. It does not replace discovery when the protein list is still open.
How to Choose Among Targeted Methods and Related Routes
Use list status, matrix risk, claim unit, and throughput need together.
Project signal |
Method path that often fits |
Recheck before committing |
|---|---|---|
Closed panel, stable transitions, high sample count |
MRM/SRM after matrix-matched lock |
Skip qualifier-ratio review |
Closed panel, interference-limited peptides |
PRM or hybrid MRM plus PRM |
Force whole panel onto weak MRM |
Absolute ng/mL or fmol required |
SIS or AQUA design plus calibrators |
Relative-only method for amount claims |
List still open after screening |
Discovery DDA or DIA first |
Targeted quote on unnamed proteins |
Biomarker or pathway remeasurement |
Locked panel version and matched prep |
Expand panel mid cohort |
Need method and QC notes |
Deliverables locked before acquisition |
Interpret tables without assay context |
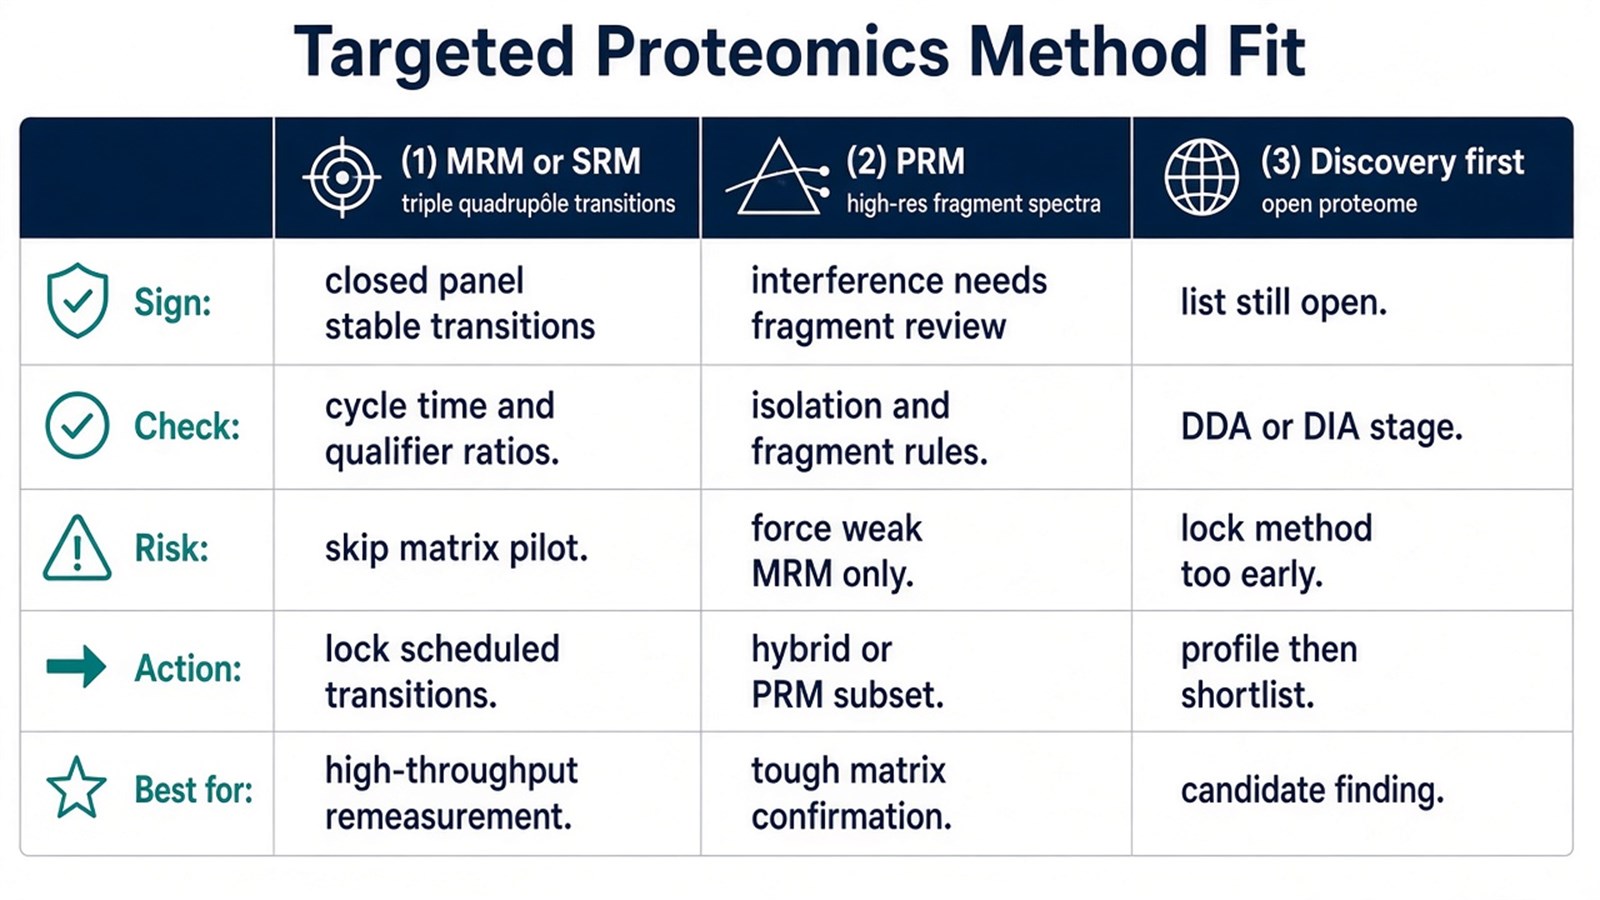
Figure 2. Choose MRM/SRM or PRM from matrix risk on a closed panel; keep discovery first when candidates are still unnamed.
Project scoping before shipment belongs on Scoping a Targeted Mass Spectrometry Project: Analyte Panel, Platform, and Validation Deliverables. Teams with closed peptide lists, matrix notes, and claim units can review MtoZ Biolabs Targeted Proteomics Service when those fields are declared in the project record.
Frequently Asked Questions
1. What is targeted proteomics in one sentence?
It is selective LC-MS/MS quantitation of predefined proteotypic peptides that represent a closed protein or peptide panel.
2. Are MRM and PRM both targeted methods?
Yes. Both remeasure predefined peptides. MRM/SRM uses scheduled transitions. PRM uses high-resolution precursor isolation and fragment spectra.
3. Is DIA or SWATH a core targeted method?
No. DIA/SWATH is discovery-class acquisition when the analyte list is not closed before the run. It may support later mining, but it is not a substitute for a locked MRM or PRM panel.
4. When is absolute quantification required?
When the decision needs concentration or amount units. Relative ratios are not concentrations without SIS or AQUA design and a declared calibration model.
5. What must be locked before calling a study targeted?
A versioned peptide list, acquisition mode, matched prep rules, claim unit, QC plan, and deliverable list.
MtoZ Biolabs can provide targeted proteomics analysis using MRM, PRM, and isotope-assisted absolute quantification for predefined protein or peptide panels, including method development and QC-supported reporting when panel design and study goals are documented.
If further evaluation of MRM versus PRM choice, relative versus absolute claim design, or discovery-to-targeted staging is needed, project details can be submitted for a feasibility review.
Reference
- V. Lange, P. Picotti, B. Domon, R. Aebersold (2008). Selected reaction monitoring for quantitative proteomics: a tutorial. Mol. Syst. Biol., 4, 222. https://doi.org/10.1038/msb.2008.61
- A.C. Peterson, J.D. Russell, A.A. Klammer, P.F. Doubleday, H. Liu, J.N. Anderson, M.J. MacCoss (2012). Parallel reaction monitoring for high resolution and high mass accuracy quantitative, targeted proteomics. Mol. Cell. Proteomics, 11, 1475-1488. https://doi.org/10.1074/mcp.O112.020271
- S. Gallien, E. Duriez, C. Crone, M. Kellmann, T. Moehring, B. Domon (2012). Targeted proteomic quantification on quadrupole-orbitrap mass spectrometer. Mol. Cell. Proteomics, 11, 1709-1723. https://doi.org/10.1074/mcp.O112.019802
- S.A. Gerber, J. Rush, O. Stemman, M.W. Kirschner, S.P. Gygi (2003). Absolute quantification of proteins and phosphoproteins from cell lysates by tandem MS. Proc. Natl. Acad. Sci. U.S.A., 100, 6940-6945. https://doi.org/10.1073/pnas.0832254100
- S.A. Carr, S.E. Abbatiello, B.L. Ackermann, C. Borchers, B. Domon, E.W. Deutsch, R.P. Grant, A.N. Hoofnagle, R. Huttenhain, J.M. Koomen, D.C. Liebler, T. Liu, B. MacLean, D.R. Mani, E. Mansfield, H. Neubert, A.G. Paulovich, L. Reiter, O. Vitek, R. Aebersold, A.J. Smith, et al. (2014). Targeted peptide measurements in biology and medicine: best practices for sample preparation, assay standardization, and assay validation. Mol. Cell. Proteomics, 13, 907-917. https://doi.org/10.1074/mcp.M113.036095
- M. Bantscheff, S. Lemeer, H.U. Haab, B. Kuster (2012). Quantitative mass spectrometry in proteomics: a critical review. Anal. Bioanal. Chem., 404, 939-965. https://doi.org/10.1007/s00216-012-5892-4
Support Documents
Protein Quantification for Deeper Discovery
Gain a clearer understanding of protein quantification approaches and their applications in studying protein abundance changes by downloading our detailed brochure.