



Multiple reaction monitoring mass spectrometry (MRM-MS) is the targeted LC-MS/MS detection mode that isolates scheduled peptide precursors, fragments them, and passes one or more predefined product ions per dwell for quantitative extraction (Lange et al. 2008). In project language, MRM-MS names the mass spectrometry layer of a closed-panel assay. Selected reaction monitoring (SRM) uses the same triple quadrupole transition logic. Parallel reaction monitoring (PRM) records a high-resolution product-ion spectrum so fragments can be reviewed after acquisition (Peterson et al. 2012). The guide below integrates MRM-MS and SRM with workflow, locked parameters, limits, and project fit. It is not a clinical oncology tour, not a metabolomics residue methods page, and not a fixed detection-limit promise.
MRM-MS peak groups support relative change of predefined peptides across frozen groups. Amount on named peptides needs stable isotope-labeled internal standard (SIS) peptides and a calibration model in the study matrix (Gerber et al. 2003). Triple quadrupole selectivity suppresses much chemical noise. It does not remove matrix suppression or create a diagnostic cutoff by itself (Bantscheff et al. 2012).
In MtoZ Biolabs targeted offerings, MRM-MS or SRM methods are built on triple quadrupole LC-MS/MS when the peptide panel is closed and matrix-matched transitions support the cohort block. Service intake appears in SRM-MRM Quantitative Proteomics Services.
What MRM-MS Detection Covers in a Targeted Record
Four detection layers belong in the brief.
Isolation scope schedules a precursor m/z window at a declared retention time. Only peptides on the inclusion list enter the first quadrupole.
Fragmentation scope uses the collision cell to generate product ions from that precursor under locked collision energy.
Analyzer scope passes one or more prechosen product ions through the third quadrupole for each dwell. Quantification uses extracted transition chromatograms, not a proteome-wide intensity matrix.
Claim scope is relative peptide change, or calibrated amount when SIS and a curve are declared. Protein inference still depends on surrogate uniqueness and digest mapping.
MRM-MS detects scheduled precursor-product pairs. It does not profile unnamed proteins from the same file.
MRM and SRM Versus PRM-MS and Versus Discovery MS
Scan mode and instrument class should be chosen separately from the project question.
MRM and SRM name the same scheduled transition pattern on triple quadrupole LC-MS/MS. The project record should lock one method name and one transition list version.
PRM-MS isolates the precursor and records a high-resolution product-ion spectrum (Gallien et al. 2012). PRM-MS fits when MRM transition ratios remain unstable after matrix-matched optimization. Route choice after matrix pilot appears in Parallel Reaction Monitoring vs MRM: Which Fits?.
Discovery data-dependent acquisition (DDA) or data-independent acquisition (DIA) mass spectrometry profiles an open or semi-open list. MRM-MS is the follow-up scan after candidates are named.
Small-molecule residue or metabolite MRM methods are a different assay class. Peptide uniqueness, digest mapping, and protein inference rules do not transfer from those libraries.
End-to-End Workflow From MRM-MS Acquisition to a Contrast Table
A practical MRM-MS spine usually runs as follows.
- Freeze contrast, replicates, and relative versus amount claim.
- Close the peptide panel and map each peptide to its protein or site.
- Match extraction, digest, and load across all contrast arms.
- Select transitions and collision energies on matrix-matched digests.
- Build dwell times and retention windows with realistic cycle time versus peak width.
- Size biological n plus quality control (QC), blanks, and SIS if used.
- Acquire interleaved study and QC injections when cohort size allows.
- Integrate transitions; review ratios and retention drift against locked rules.
- Test predefined contrast only; archive transition list, instrument method, and software with deliverables.
Steps 1 through 3 protect the sample record. Steps 4 through 9 are the MRM-MS detection and informatics record. Sample and pretreatment scoping appears in Scoping an MRM Quantitation Project: Transition Design, Sample Count, and Validation Deliverables.
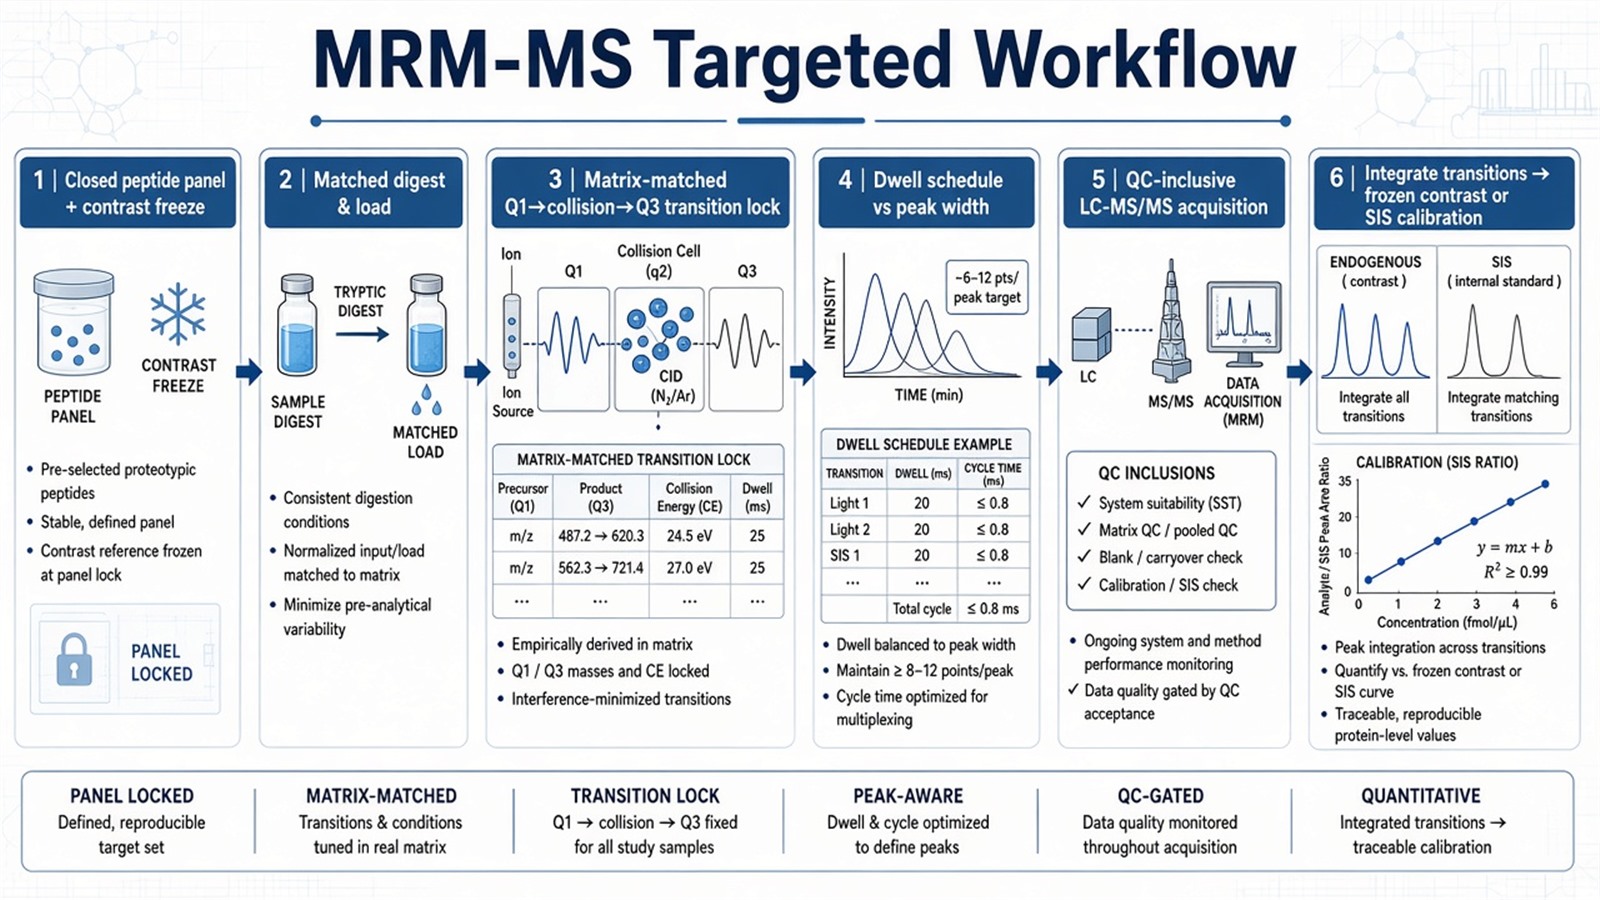
Figure 1. MRM-MS comparability depends on a closed transition list and matched pretreatment before group contrasts are tested.
Parameters That Must Be Locked in MRM-MS Methods
Layer |
Parameter that must be declared |
Failure mode if ignored |
|---|---|---|
Naming |
MRM or SRM locked to one list version |
Two names for the same method file |
Panel |
Peptide list and protein or site mapping |
Moving targets mid cohort |
Transitions |
Precursor, products, dwell, collision energy |
Neat-standard methods fail in matrix |
Schedule |
Cycle time versus chromatographic peak width |
Undersampled or missed peaks |
Prep |
Matched extraction and digest |
Peptide-pool shift as biology |
Load |
Mass on column from pilot |
Saturation or dropped transitions |
SIS |
Spike step aligned to claim |
Relative peaks read as concentration |
QC |
Pools, blanks, and bracket pattern |
Undetected batch drift |
Contrast |
Frozen before peak review |
Post hoc arm relabeling |
Deliverable |
Table, transition list, method notes |
Report rebuilt after acquisition |
Limits follow the same table. MRM-MS does not remove ion suppression in plasma or tissue. Large transition counts do not raise throughput if cycle time exceeds peak width. High sensitivity language does not replace matrix-matched pilots. Clinical diagnosis, disease progression, or personalized-medicine language needs separate validation design beyond the quantitative table.
How to Choose MRM-MS Versus Other Mass Spectrometry Routes
Use list status, matrix pilot, claim unit, and instrument context together.
Project need |
Route that often fits |
Recheck before committing |
|---|---|---|
Closed peptides, stable transitions in matrix |
MRM-MS or SRM on triple quadrupole |
Open discovery MS as the targeted table |
Transition ratios unstable after prep lock |
PRM-MS subset or hybrid panel |
Whole-panel PRM without interference evidence |
Open proteome relative profiling |
Discovery DDA or DIA first |
MRM-MS on unnamed proteins |
Amount on named peptides |
MRM-MS or PRM-MS with SIS and curve |
Relative peaks as concentration |
Transferred MRM method, same matrix |
Acquisition plus QC |
Assume free method rebuild |
Residue or metabolite question |
Separate small-molecule MRM assay |
Peptide protein-inference rules |
Biomarker or clinical cutoff intent |
Targeted panel plus validation design |
MRM-MS table as diagnostic test |
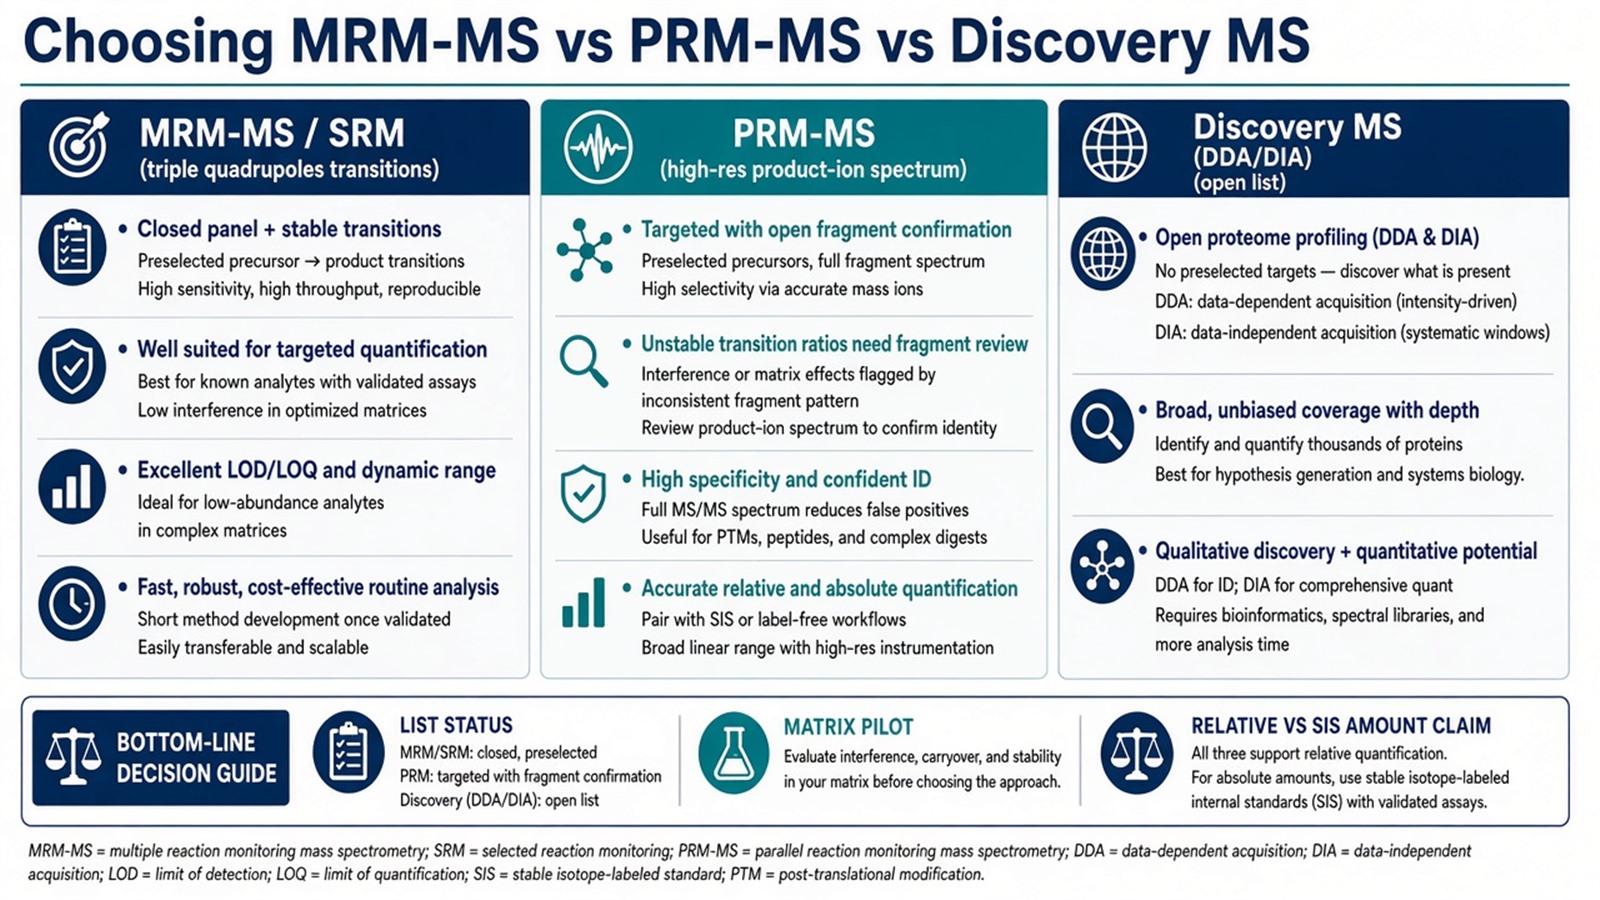
Figure 2. Choose MRM-MS when a closed transition list is stable in the study matrix; use PRM-MS or discovery MS when list status or interference changes the need.
Teams with closed peptide targets, instrument class notes, and contrast maps can review MtoZ Biolabs SRM-MRM Quantitative Proteomics Services when those fields are declared in the project record.
Frequently Asked Questions
1. Is MRM-MS the same as MRM or SRM?
Yes for most targeted proteomics briefs. MRM-MS emphasizes the mass spectrometry detection layer. SRM and MRM name the same scheduled transition pattern. The assay still needs a closed panel, matched pretreatment, and a frozen contrast.
2. Does MRM-MS work without internal standards?
Relative contrasts can run light-only when that claim is declared. SIS and a curve become necessary when amount units are required. Triple quadrupole selectivity does not replace calibration.
3. Can MRM-MS quantify many proteins without a closed list?
No. Multiple names concurrent scheduled transitions. Proteins not on the inclusion list are not quantified from that method file.
4. Does MRM-MS cover clinical biomarker diagnosis on this page?
MRM-MS can remeasure closed peptide candidates. Diagnostic or prognostic cutoffs need separate validation design beyond the quantitative table.
5. When should projects move from MRM-MS to PRM-MS?
When matrix-matched pilots still show unstable transition ratios after cleanup and collision-energy work, a PRM-MS subset can be scoped on the same chromatography family when possible.
MtoZ Biolabs can discuss MRM-MS scoping when peptide panel, transition method, and contrast design are documented.
If further evaluation of MRM-MS versus PRM-MS, SIS calibration, or discovery-to-targeted handoff is needed, project details can be submitted for a feasibility review.
Reference
- V. Lange, P. Picotti, B. Domon, R. Aebersold (2008). Selected reaction monitoring for quantitative proteomics: a tutorial. Mol. Syst. Biol., 4, 222. https://doi.org/10.1038/msb.2008.61
- A.C. Peterson, J.D. Russell, A.A. Klammer, P.F. Doubleday, H. Liu, J.N. Anderson, M.J. MacCoss (2012). Parallel reaction monitoring for high resolution and high mass accuracy quantitative, targeted proteomics. Mol. Cell. Proteomics, 11, 1475-1488. https://doi.org/10.1074/mcp.O112.020271
- S.A. Gerber, J. Rush, O. Stemman, M.W. Kirschner, S.P. Gygi (2003). Absolute quantification of proteins and phosphoproteins from cell lysates by tandem MS. Proc. Natl. Acad. Sci. U.S.A., 100, 6940-6945. https://doi.org/10.1073/pnas.0832254100
- M. Bantscheff, S. Lemeer, H.U. Haab, B. Kuster (2012). Quantitative mass spectrometry in proteomics: a critical review. Anal. Bioanal. Chem., 404, 939-965. https://doi.org/10.1007/s00216-012-5892-4
- S. Gallien, E. Duriez, A. Crone, M. Kellmann, T. Moehring, T. Domon (2012). Targeted proteomic quantification on quadrupole-orbitrap mass spectrometers. Mol. Cell. Proteomics, 11, 1709-1723. https://doi.org/10.1074/mcp.O112.019232
