



An immunoprecipitation mass spectrometry protocol, or IP-MS protocol, asks which proteins remain with an endogenous bait after antibody capture, Input and IgG contrast, and LC-MS/MS identification. The protocol is usable only when the antibody recovers the native bait, the control arms share one recipe, and bait enrichment is documented before any partner is ranked. It fits cells or tissues when an IP-competent antibody exists and the claim is co-recovery, not binary direct binding (Hein et al. 2015). Prefer this route when antibody performance can be shown in a pilot. Reassess when the antibody is Western-only, Input and IgG are omitted, or pathway enrichment is used to repair a missing bait.
Researchers who already have a named bait, an antibody, and a written interaction claim can also review the MtoZ Biolabs IP-Mass Spectrometry Service and Protein-Protein Interaction Analysis for project-specific IP-MS protocol feasibility.
What an Immunoprecipitation Mass Spectrometry Protocol Must Answer
A named IP-MS protocol is not RIPA lysis, overnight antibody binding, trypsin, and a network figure. Those steps can appear in a notebook. They do not make the identification table a partner list. The protocol has to answer one sentence: which proteins are enriched with this endogenous bait versus matched Input and IgG after LC-MS/MS.
Co-IP-MS is the same capture class when the written claim is open partner discovery. IP-MS is the right first name when bait recovery itself must be shown. Neither name authorizes presence-or-absence calling. Capture-to-instrument layers after these gates are locked are covered in IP-MS Protocol: From Antibody Capture to LC-MS/MS. How the wet-lab arms stay comparable after elution is covered in How to Prepare Immunoprecipitation Samples for Mass Spectrometry.
The protocol fits cultured cells or tissues when a protein reference database exists and an IP-competent antibody is supplied by the study. It is a poor first protocol when the antibody only works on a denatured blot, when tagging is required because no IP reagent exists, or when the claim is reconstituted binding of a purified domain.
An immunoprecipitation mass spectrometry protocol supports candidate co-recovery under stated controls. It does not convert a search report into proof of a contact.
Antibody Specificity Before the First Incubation
Antibody specificity is the first protocol line, not a catalog footnote. The capturing reagent must recover the intended native bait from the planned lysate without a dominant off-target that then occupies the MS table. Application-specific validation remains necessary because Western performance does not establish IP performance (Uhlen et al. 2016). How to qualify that reagent is covered in How to Choose the Right Antibody for Co-IP Assays?.
Record clone, lot, host species, isotype, and protein A or protein G pairing before any overnight incubation is copied from another bait. Do not write overnight binding as a required step. Incubation time is titrated against bait recovery versus Input, not against a transferred recipe. Extra antibody usually increases chain peptides faster than it increases bait peptides.
If no IP-competent antibody exists, the next protocol is tagged AP-MS or a pull-down, not a longer LC-MS/MS run. That switch is compared in AP-MS vs IP-MS: How to Choose the Right Interactome Workflow.
Input and IgG Belong in the Protocol, Not in an Appendix
Input and IgG are protocol arms. They are not optional lanes added after a pathway map is drawn.
Input is taken before capture. It shows whether the bait was present and extractable, and it provides a baseline intensity for enrichment. If Input lacks the bait, a missing IP-MS identification is an expression or extraction problem.
IgG or bead controls must use the same lysate batch, lysis, incubation, wash, elution, digestion, and search settings as the bait IP. Preclearing can lower some sticky binders. It does not replace IgG. Empty-tag controls from AP-MS do not replace IgG in an antibody IP-MS protocol. Asymmetric wash or elution between bait IP and IgG makes every later score unreadable.
Protocol arm |
Design job |
What it cannot do |
|---|---|---|
IP-competent antibody |
Recover the native bait without a dominant off-target |
Convert a Western-only clone into an IP-MS handle |
Input |
Show starting bait and a baseline for enrichment |
Prove the antibody works |
IgG or bead |
Estimate handle-driven background under the same recipe |
Replace Input, or replace IgG with preclearing |
Matched digestion and search |
Keep bait IP and IgG on one identification scale |
Repair a failed bait IP with a longer gradient |
Replicates |
Stabilize the same protocol across biological arms |
Merge unlike lysis or wash chemistries |
A practical protocol sequence is:
- Write the co-recovery claim the identification table must support.
- Confirm native bait recovery versus Input with the chosen antibody.
- Process matched IgG with the same wash and elution.
- Digest and search bait IP, Input, and IgG together.
- Rank remaining names only after bait enrichment is shown.
Generic Co-IP design of those arms is covered in Designing an Efficient Co-immunoprecipitation (Co-IP) Protocol.
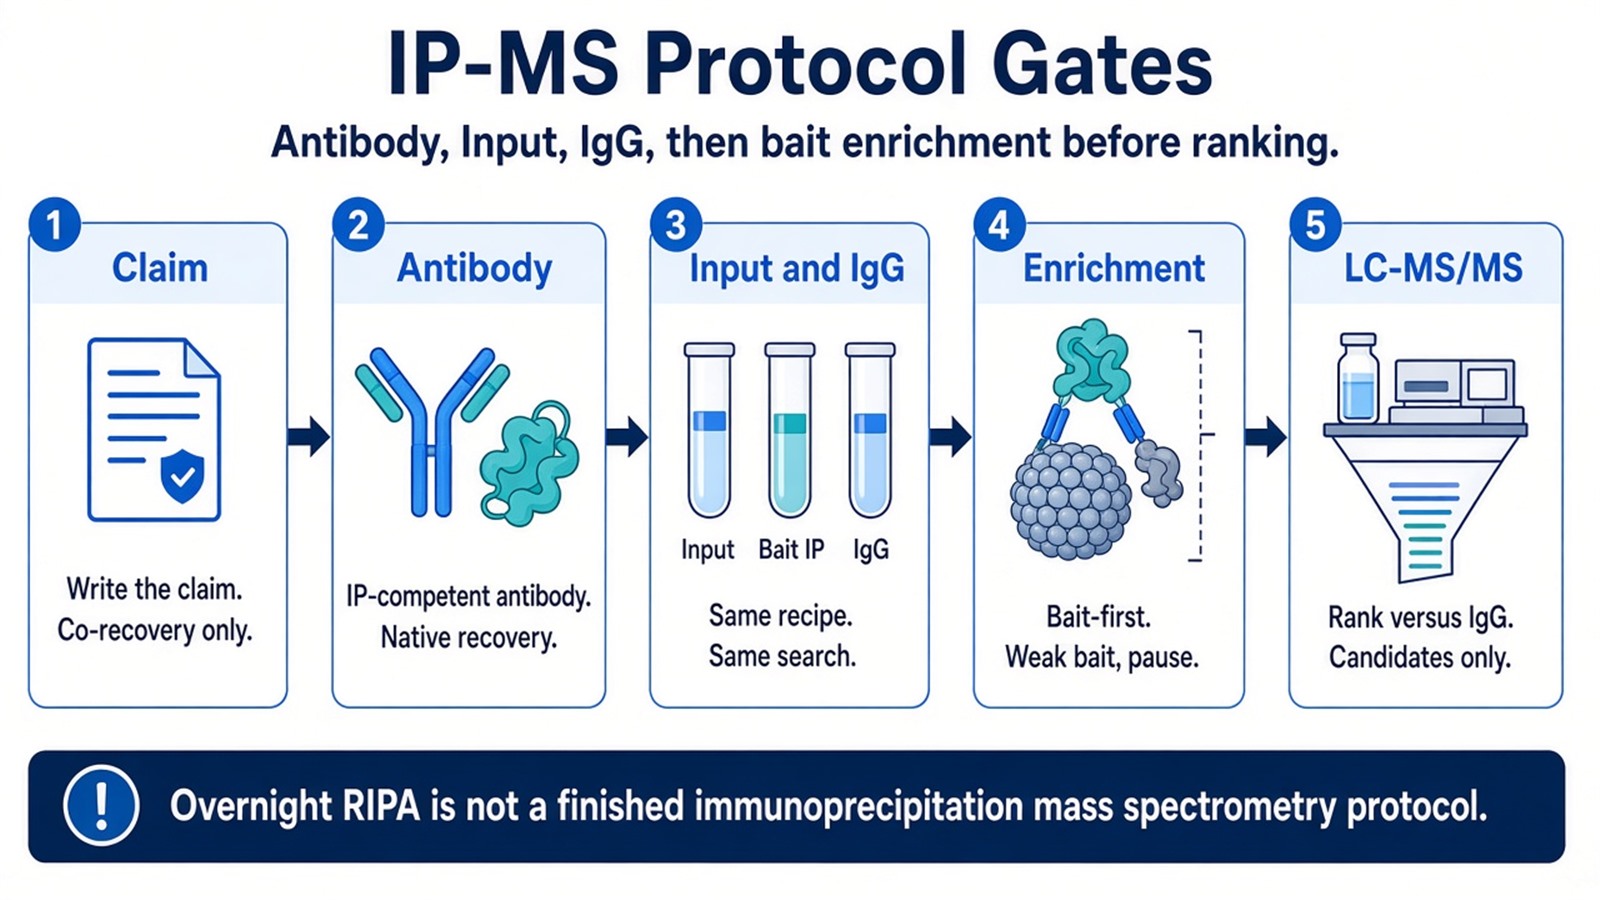
Figure 1. An IP-MS protocol starts with the antibody and the control arms. RIPA plus overnight incubation is not a finished method.
Enrichment Efficiency and Mass Spectrometry Credibility
Enrichment efficiency is bait-first. If the bait is weak versus Input, a long protein table is not a stronger protocol. Relative intensity can compare the same protein across Input, IgG, and bait IP. It does not rank unrelated proteins as tighter binders. Identification depth in a Co-IP-style eluate is typically hundreds to about a thousand proteins, depending on the sample. Depth without bait recovery is background.
Database searching identifies peptide-spectrum matches. It does not filter handle background. Network or functional enrichment analysis can annotate a shortlist. It cannot invent bait recovery or replace IgG. Treating pathway software as the last protocol step is the usual reason an unfiltered IP-MS table is over-read.
Identification of a protein in the eluate is not a confidence call. Ranking requires bait recovery, Input-aware enrichment, and a declared contrast against IgG. Probabilistic scoring such as SAINT uses bait versus control counts to assign a probability that a prey is specifically co-purified (Choi et al. 2011). A contaminant catalog such as the CRAPome is a prior on control frequency (Mellacheruvu et al. 2013). Neither score repairs a Western-only antibody.
After those gates, a protein-level value can support: protein B is enriched with bait A versus matched IgG inside this immunoprecipitation mass spectrometry protocol. Peptide support, antibody clone, and wash chemistry belong next to that statement.
If the bait itself is missing, stop ranking and diagnose recovery, as in Why Was the Target Protein Not Detected by IP-MS?. When the next decision is open MS versus a named-pair blot, see IP-MS vs Co-IP Western Blot: Which Method Fits Your Study?.
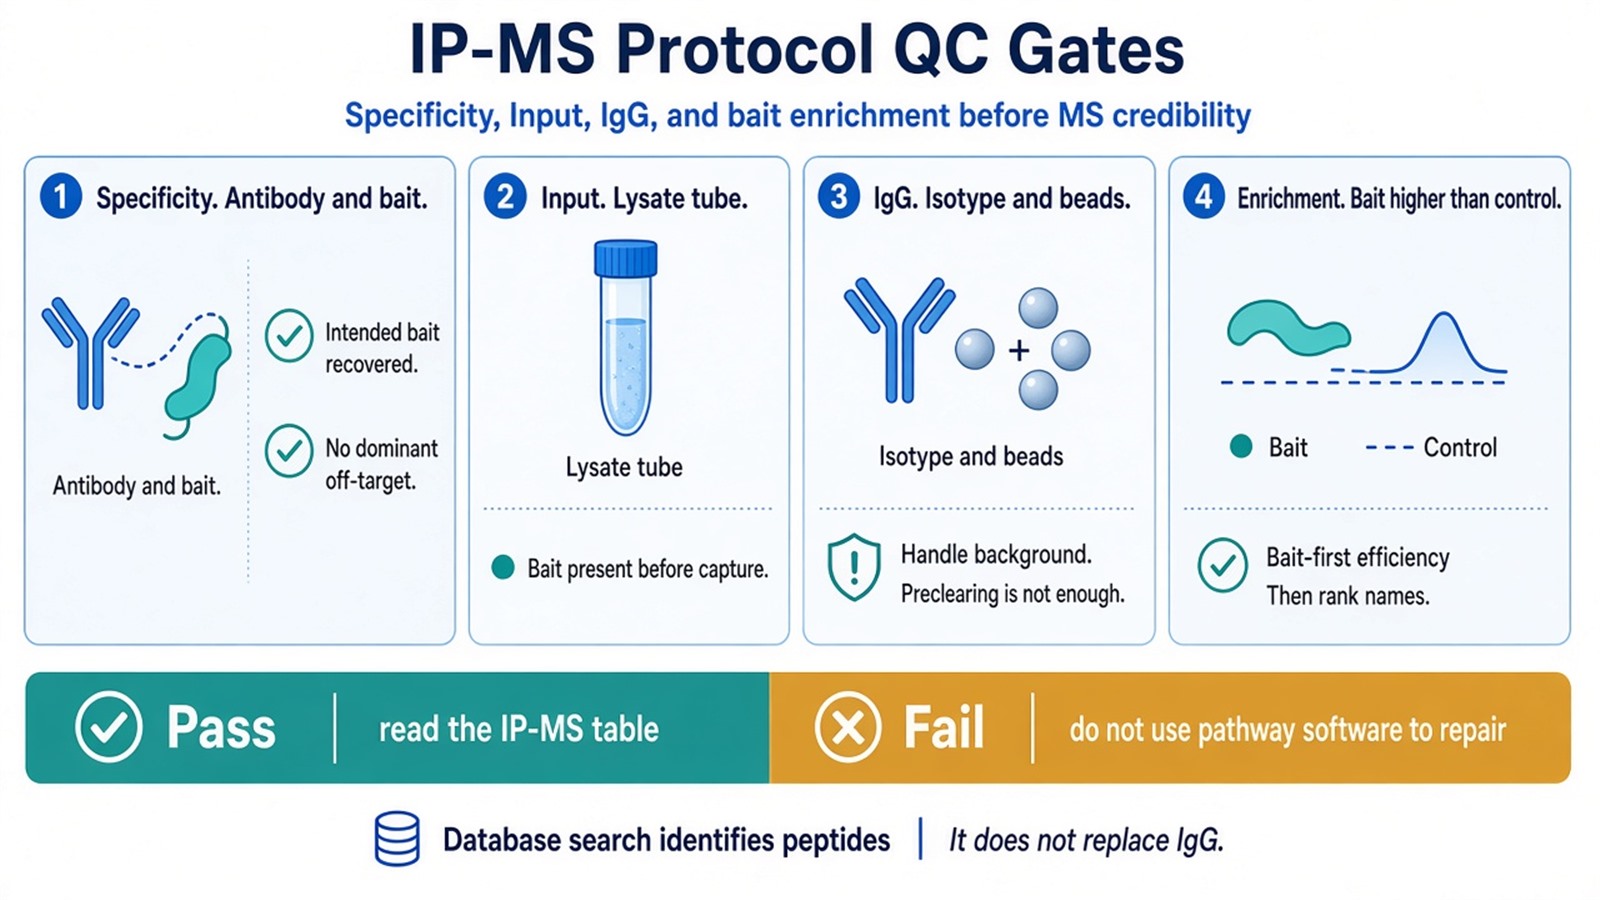
Figure 2. Mass spectrometry credibility is a protocol outcome. Pathway enrichment does not replace bait recovery versus Input and IgG.
Common Biases and Result Boundaries
The common bias is copying a generic IP-MS protocol, then reading every identified name as an interactor. Another bias is using RIPA or another harsh mix because it looks clean on blot, then concluding that missing partners mean no interaction. Under-washing creates the opposite error: bait IP and IgG look similar, so subtraction makes every name look specific. Overnight incubation does not correct a Western-only antibody.
Do not write a successful bait identification as proof that every remaining protein contacts the bait. Do not write a failed IP-MS protocol as proof that no interaction exists in the cell. Absence can mean the wrong antibody, over-washing, failed digestion, or a bait that is not expressed in the tested state.
Prefer this immunoprecipitation mass spectrometry protocol when the endogenous bait can be recovered and the claim is candidate co-recovery. Prefer AP-MS or GST pull-down when no IP-competent antibody exists. Prefer a targeted Co-IP blot when only a named pair matters and discovery depth is unnecessary. Screening-level maps are covered in Measuring Protein-Protein Interactions. Mixed enrichment-MS projects can be reviewed under the MS-Based Protein-Protein Interaction Analysis Service.
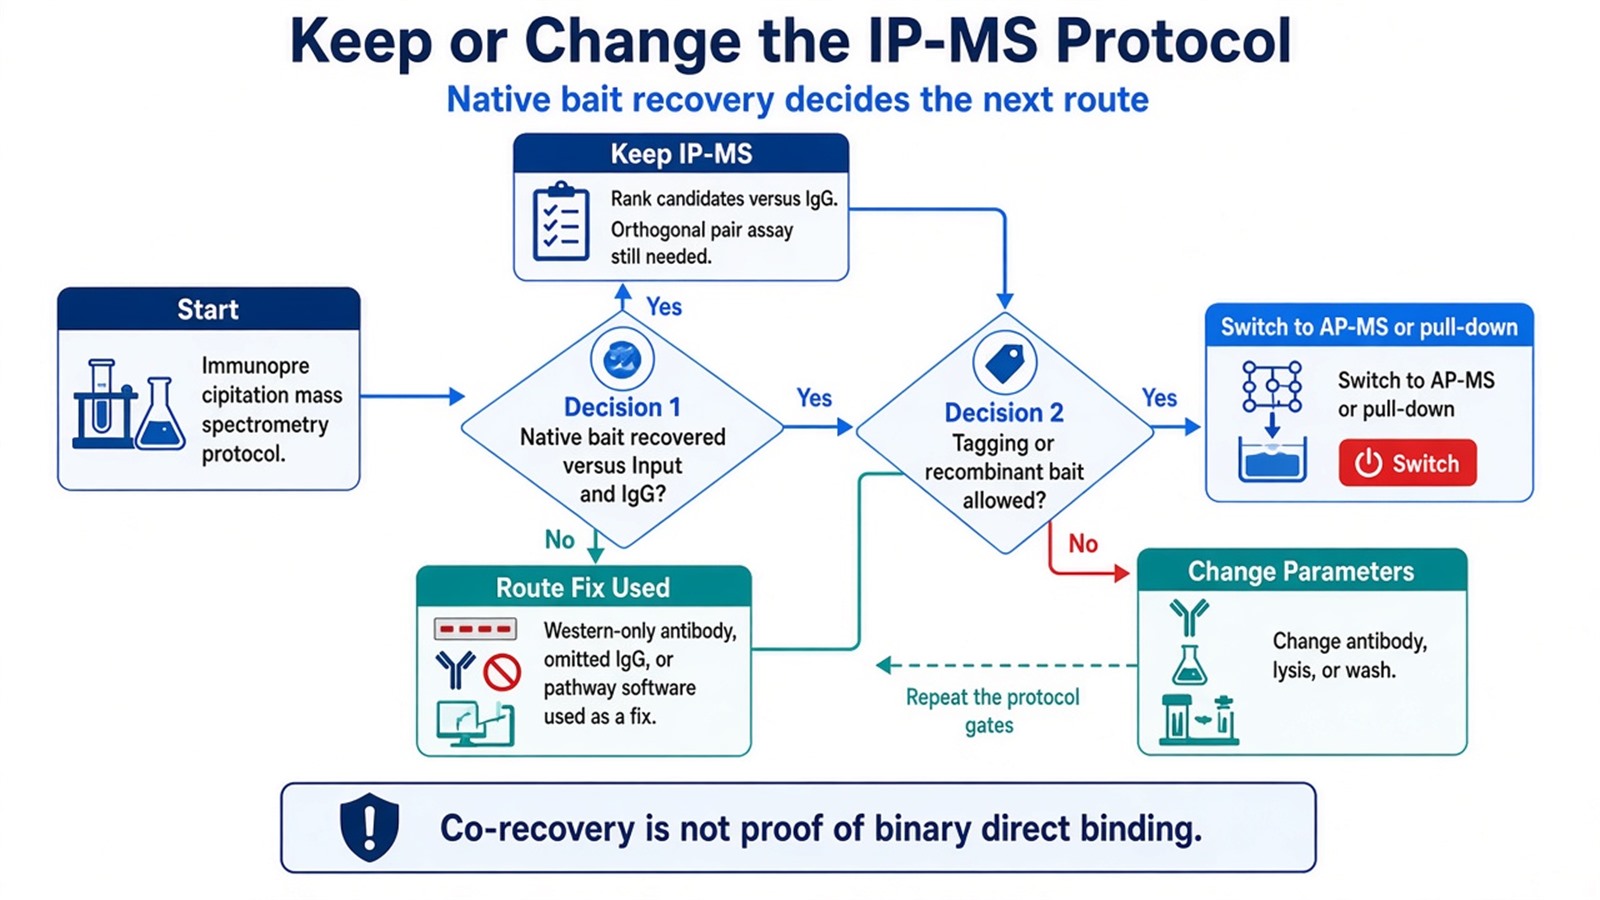
Figure 3. Stay with the IP-MS protocol only while native bait recovery is documented. A copied overnight recipe is not a completed design.
An immunoprecipitation mass spectrometry protocol supports candidate co-recovery after antibody, Input, IgG, and bait enrichment are shown. It does not certify a binary interactome.
Once antibody status, Input and IgG plans, enrichment checks, and the expected identification output are defined, the project can be scoped as an IP-MS study. Researchers can review the MtoZ Biolabs IP-Mass Spectrometry Service for sample evaluation and protocol planning.
Frequently Asked Questions
1. Is overnight incubation required in an IP-MS protocol?
No. Incubation is titrated against bait recovery versus Input. A copied overnight step does not fix a Western-only antibody.
2. Can RIPA be used as the default lysis in every IP-MS protocol?
Only if that mix recovers the native bait with acceptable partner retention. A blot-clean lysate can still strip the associations the claim needed.
3. Does database search plus pathway analysis complete the protocol?
No. Search identifies peptides. Pathway software annotates a shortlist. Neither replaces Input, IgG, or bait enrichment.
4. Can preclearing replace an IgG control?
No. Preclearing can lower some bead binders. IgG still estimates handle-driven background under the same recipe.
5. When should this protocol change to AP-MS or pull-down?
Change when no IP-competent antibody recovers the native bait and tagging or a recombinant bait is acceptable. That is a different handle, not a longer MS method.
Reference
- M. Uhlen, A. Bandrowski, S. Carr, A. Edwards, J. Ellenberg, E. Lundberg, D. L. Rimm, H. Rodriguez, T. Hiltke, M. Snyder, T. Yamamoto (2016). A proposal for validation of antibodies. Nat. Methods, 13, 823-827. https://doi.org/10.1038/nmeth.3995
- M.Y. Hein, N.C. Hubner, I. Poser, J. Cox, N. Nagaraj, Y. Toyoda, I.A. Gak, I. Weisswange, J. Mansfeld, F. Buchholz, et al. (2015). A human interactome in three quantitative dimensions organized by stoichiometries and abundances. Cell, 163, 712-723. https://doi.org/10.1016/j.cell.2015.09.053
- H. Choi, B. Larsen, Z.Y. Lin, A. Breitkreutz, D. Mellacheruvu, D. Fermin, Z.S. Qin, M. Tyers, A.C. Gingras, A.I. Nesvizhskii (2011). SAINT: probabilistic scoring of affinity purification-mass spectrometry data. Nat. Methods, 8, 70-73. https://doi.org/10.1038/nmeth.1541
- D. Mellacheruvu, Z. Wright, A.L. Couzens, J.P. Lambert, N.A. St-Denis, T. Li, Y.V. Miteva, S. Hauri, M.E. Sardiu, T.Y. Low, et al. (2013). The CRAPome: a contaminant repository for affinity purification-mass spectrometry data. Nat. Methods, 10, 730-736. https://doi.org/10.1038/nmeth.2557
