



Current-state challenges in targeted proteomics are most useful when treated as troubleshooting problems—not as a trends brochure about clinical translation or artificial intelligence. Targeted proteomics schedules closed proteotypic peptides by selected reaction monitoring (SRM), multiple reaction monitoring (MRM), or parallel reaction monitoring (PRM) on liquid chromatography-tandem mass spectrometry (LC-MS/MS) (Lange et al. 2008; Peterson et al. 2012). In project language, today’s recurring failures cluster around peptide design, reference-standard (SIS) readiness, acquisition or data-review overload, and choosing targeted work while the list is still open. The guide below maps those challenges into a diagnostic workflow, locked parameters, limits, and recovery fit. It is not a vaccine or companion-diagnostics catalog, not a GLP/GCP compliance claim, and not a forecast that DIA-PRM or automation removes method design.
Unmatched prep and untested surrogates still break quantitative claims (Bantscheff et al. 2012). Relative fold change and amount with stable isotope-labeled internal standard (SIS) peptides remain separate claim units (Gerber et al. 2003).
In MtoZ Biolabs targeted offerings, challenge review starts after symptom class, panel status, matrix, and claim unit are written in the brief. Service intake appears in Targeted Proteomics Service.
What Challenge Layers Must Be Separated
Four challenge layers belong in the troubleshooting record.
Peptide-design layer: poor proteotypic choice, modification-prone sites, weak ionization, or uniqueness failures that look like “low sensitivity.”
SIS-and-claim layer: missing, mismatched, or wrongly spiked labeled standards when amount units were promised; relative methods blamed for lacking concentration.
Acquisition-and-review layer: oversized panels, unstable transitions or windows, and PRM file burden without locked extraction rules.
Route-mismatch layer: forcing targeted build while candidates are still open, or rediscovering a closed panel every batch.
Name the challenge layer before changing platform, software, or “next-generation” workflow labels.
Instrument-level transition, cycle-time, and matrix interference recovery appear in Targeted Mass Spectrometry Assay Failing? Troubleshooting Transitions, Cycle Time, and Matrix Interference. This page stays on project-level challenges behind those symptoms.
How Today’s Challenges Show Up in Practice
Peptide design challenges appear when discovery IDs do not survive matrix-matched targeted pilots. Rebuild surrogates before blaming the mass spectrometer.
SIS challenges appear when absolute language is requested without qualified labeled peptides, spike-step declaration, or matrix-matched calibrators. Keep relative claims until those pieces exist.
Data-processing challenges appear when PRM full spectra or large MRM panels are reviewed without locked integration, ion-ratio, and QC rules. Software helps; it does not replace those locks.
Route-mismatch challenges appear when teams expect targeted methods to discover unnamed biology, or expect discovery alone to deliver assay-level remonitoring. MRM versus PRM choice for interference appears in Parallel Reaction Monitoring vs MRM: Which Fits?. When targeted should replace rediscovery appears in When Targeted Mass Spectrometry Outperforms Discovery.
End-to-End Troubleshooting Workflow for Current Challenges
A practical challenge-recovery spine usually runs as follows.
- Capture the failing claim: relative panel, absolute concentration, or discovery-to-targeted hand-off.
- Classify the dominant layer: peptide design, SIS/claim, acquisition/review, or route mismatch.
- Re-check proteotypic peptides in matrix-matched digests; retire weak or non-unique surrogates.
- Align claim unit with SIS readiness; drop absolute language if calibrators are not locked.
- Resize or stage the panel so cycle time and review rules can hold.
- Lock MRM transitions or PRM extraction settings; change one factor set at a time.
- Run a small pilot with declared QC; stop before full cohort if ratios or missingness fail.
- Only then consider hybrid discovery-to-targeted staging or PRM subsets for interference-limited peptides.
- Archive revised peptide list, method, and challenge notes with the deliverable.
Steps 1 through 2 diagnose. Steps 3 through 9 recover without substituting trend slogans for method locks.
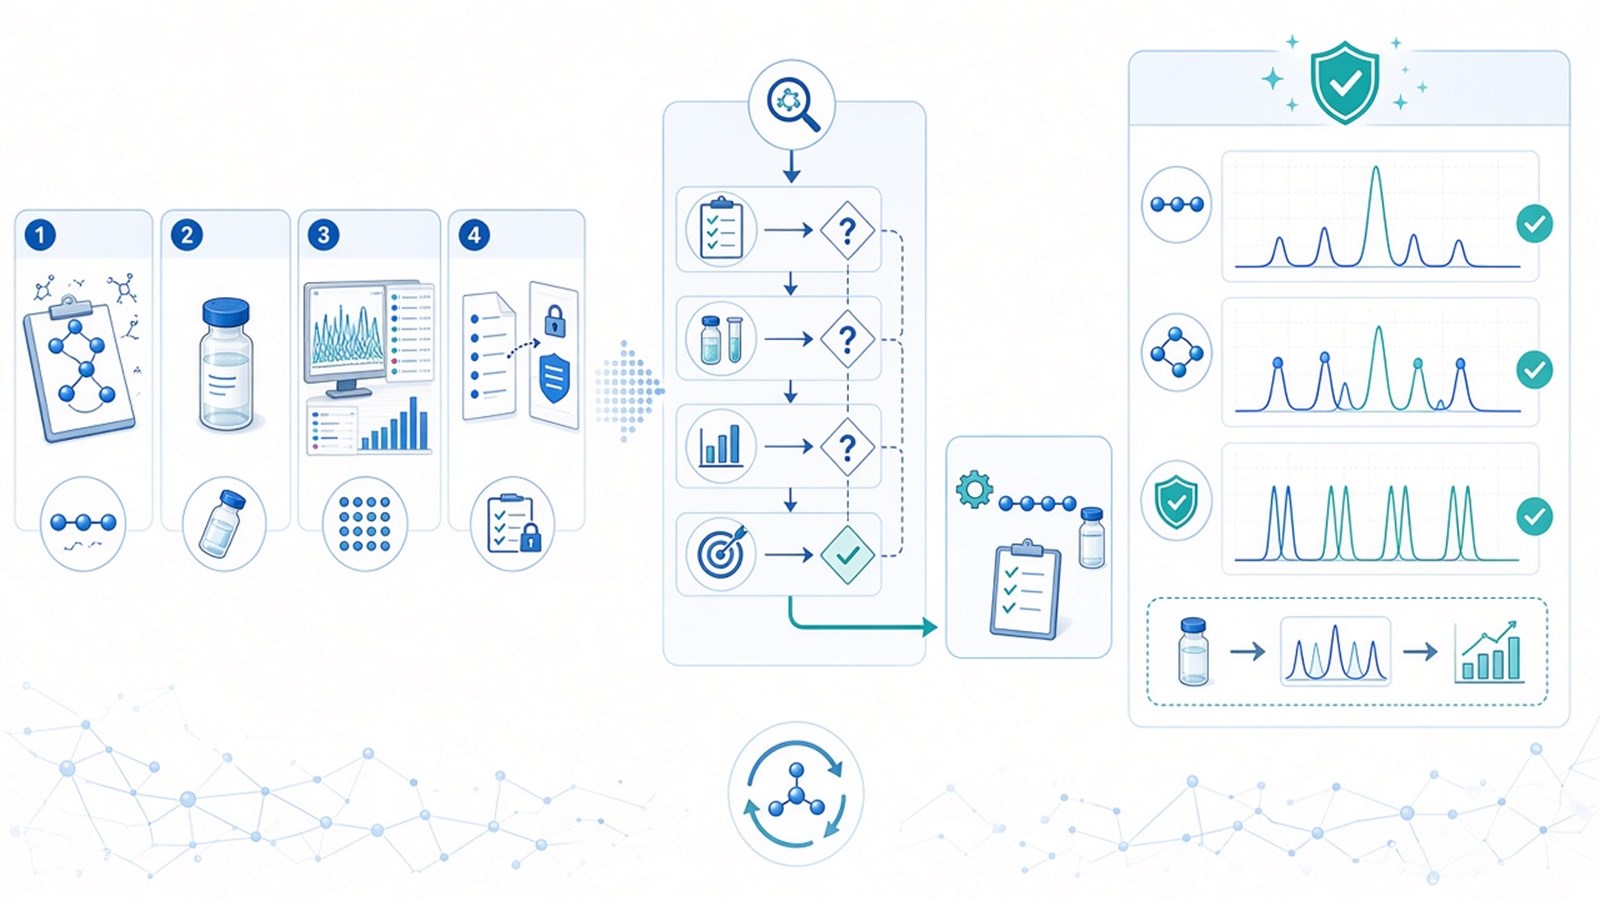
Figure 1. Treat current targeted-proteomics challenges as layered failures: design, SIS claim, acquisition review, then route mismatch.
Parameters to Re-Lock When Challenges Appear
Layer |
Parameter to re-check |
Failure mode if skipped |
|---|---|---|
Claim |
Relative vs SIS absolute |
Concentration language without calibrators |
Peptides |
Uniqueness, mod risk, matrix detectability |
Discovery ID ≠ targeted analyte |
SIS |
Sequence, spike step, documentation |
Absolute claim without standards |
Panel size |
Priority tiers vs cycle time |
Review and sampling collapse |
Platform |
MRM vs PRM rationale |
Interference unread |
Extraction |
Integration and ratio rules |
Software-dependent irreproducibility |
QC |
Blanks, pools, failed-target flags |
Undiagnosed drift |
Prep |
Matched digest and load |
Prep variance as “platform challenge” |
Pilot |
Small matrix set before cohort |
Full re-run of a broken design |
Hand-off |
Discovery list version |
Targeted build on moving targets |
Limits follow the same table. Automation, AI, or DIA hybrids do not remove peptide and matrix pilots. Regulatory or clinical diagnostic language needs a separate quality system beyond the research troubleshooting table. Hard “hundreds of targets in one run” claims are not universal capacity guarantees.
How to Choose the Recovery Path
Use challenge class, list status, and claim unit together.
Observed challenge |
Recovery path that often fits |
Recheck before committing |
|---|---|---|
Surrogates fail in matrix |
Peptide redesign + matrix pilots |
Instrument brand swap first |
Absolute asked, no SIS/curves |
Relative claim or build calibrators |
Keep absolute wording |
Panel too large to review or sample |
Stage tier-one targeted method |
Add more targets |
Unstable MRM ratios |
Alternate products or PRM subset |
Force full MRM panel |
List still open |
Discovery first |
Premature targeted build |
Closed list rediscovered each batch |
Targeted remonitoring |
Repeat open discovery |
PRM files unmanageable |
Lock extraction rules; cut panel |
New software only |
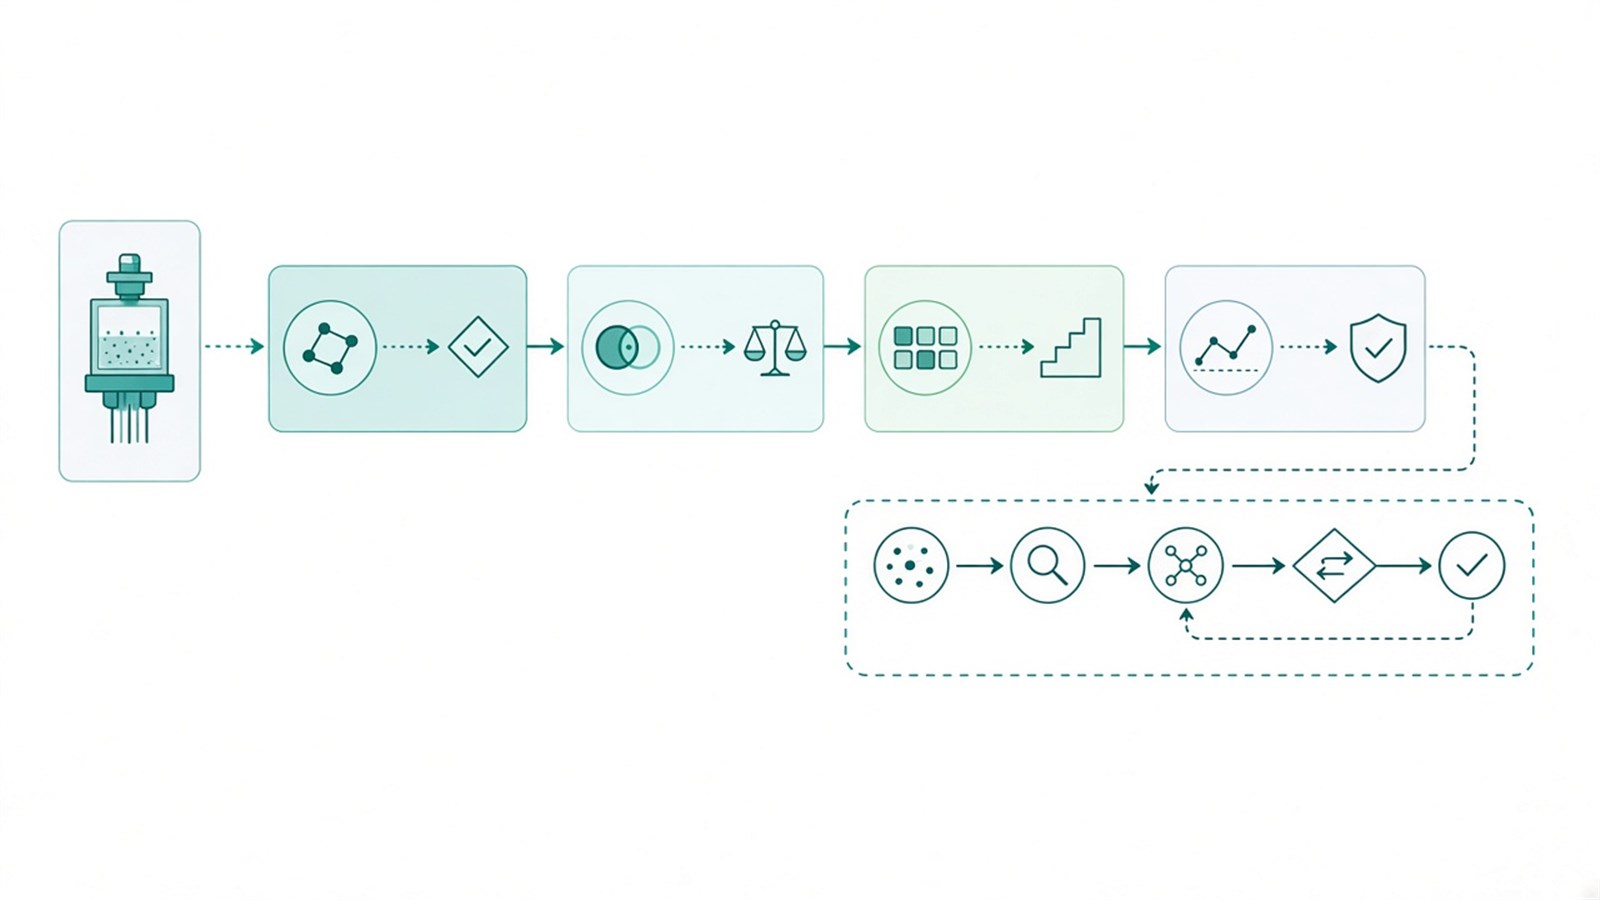
Figure 2. Match recovery to the challenge layer; do not treat trends language as a substitute for locked peptides and claims.
Teams with failing panels, SIS status, and matrix notes can review MtoZ Biolabs targeted entries when those fields are declared in the recovery record. MRM service framing appears in SRM-MRM Quantitative Proteomics Services.
Frequently Asked Questions
1. Are “development trends” a substitute for troubleshooting?
No. Hybrid DIA-targeted ideas or automation may help later. Current failures still need peptide, SIS, acquisition, and route diagnosis first.
2. Why do discovery candidates fail in targeted assays?
Because proteotypic performance, uniqueness, and matrix behavior were not locked. Targeted methods measure declared peptides, not nomination lists.
3. Is SIS always required for modern targeted proteomics?
No. SIS is required when amount units are claimed. Relative closed-panel work can proceed without absolute calibrators when that claim is declared.
4. Does PRM remove data-processing challenges?
No. Full fragment spectra can improve specificity and still need locked extraction, QC, and panel sizing.
5. When should projects leave the targeted lane?
When the protein list is open again, or when surrogates cannot be made matrix-compatible after redesign pilots.
MtoZ Biolabs can discuss targeted-proteomics challenge recovery when symptom class, panel status, and claim unit are documented.
If further evaluation of peptide redesign, SIS alignment, or MRM-to-PRM fallback is needed, project details can be submitted for a feasibility review.
Reference
- V. Lange, P. Picotti, B. Domon, R. Aebersold (2008). Selected reaction monitoring for quantitative proteomics: a tutorial. Mol. Syst. Biol., 4, 222. https://doi.org/10.1038/msb.2008.61
- A.C. Peterson, J.D. Russell, A.A. Klammer, P.F. Doubleday, H. Liu, J.N. Anderson, M.J. MacCoss (2012). Parallel reaction monitoring for high resolution and high mass accuracy quantitative, targeted proteomics. Mol. Cell. Proteomics, 11, 1475-1488. https://doi.org/10.1074/mcp.O112.020271
- S.A. Gerber, J. Rush, O. Stemman, M.W. Kirschner, S.P. Gygi (2003). Absolute quantification of proteins and phosphoproteins from cell lysates by tandem MS. Proc. Natl. Acad. Sci. U.S.A., 100, 6940-6945. https://doi.org/10.1073/pnas.0832254100
- M. Bantscheff, S. Lemeer, H.U. Haab, B. Kuster (2012). Quantitative mass spectrometry in proteomics: a critical review. Anal. Bioanal. Chem., 404, 939-965. https://doi.org/10.1007/s00216-012-5892-4
