



Quantitative proteomics methods fall into labeled and label-free families that mostly report relative abundance of the same protein across samples. Labeled routes include metabolic labeling such as SILAC and chemical isobaric tagging such as tandem mass tags (TMT) or iTRAQ, which mix channels and read isotope pairs or reporter ions. Label-free quantification (LFQ) keeps each digest as an independent liquid chromatography-tandem mass spectrometry (LC-MS/MS) run and rebuilds comparability by alignment and normalization after DDA or DIA acquisition. Absolute concentration is a different claim and usually needs stable isotope-labeled standards (SIS) with a calibration model, not ordinary discovery labeling or LFQ intensities alone. Method choice follows sample chemistry, enrollment rules, and the quantitative claim, not a single ranking of accuracy.
Researchers who already have defined samples, comparison groups, or a specific project objective can also review the MtoZ Biolabs TMT Quantitative Proteomics Analysis Service and Label-Free Quantitative Proteomics Service for project-specific feasibility and analysis planning.
Method Map: Labeled Versus Label-Free
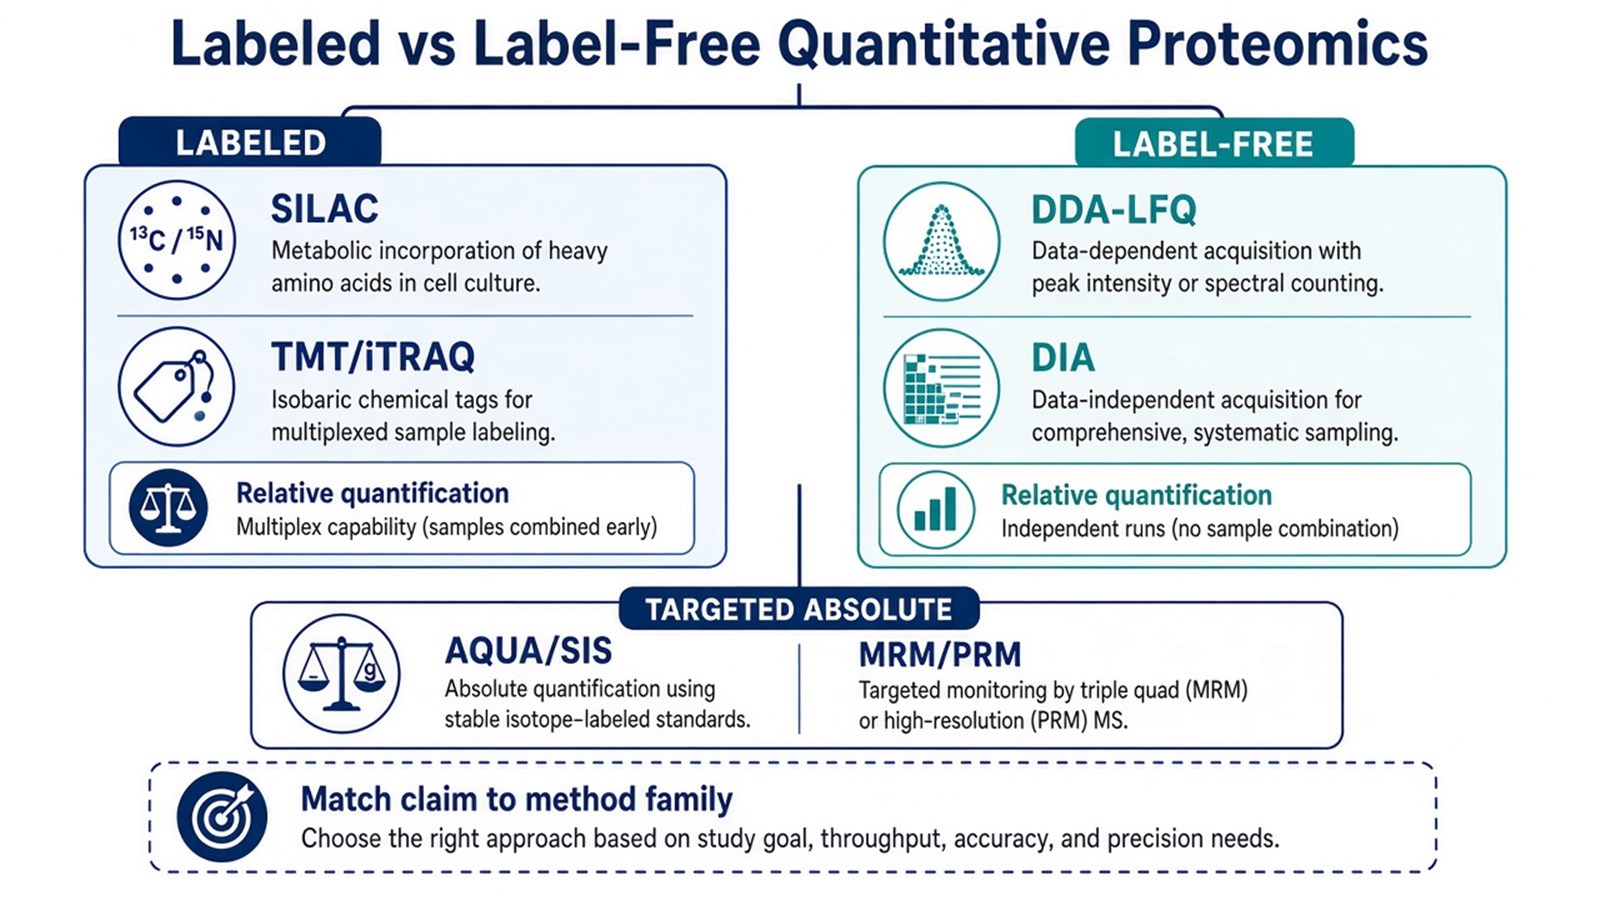
Figure 1. Labeled and label-free discovery methods answer relative questions; absolute units usually require a separate SIS-targeted design.
Method family |
Typical examples |
Core idea |
Default claim |
|---|---|---|---|
Metabolic labeling |
SILAC |
Incorporate heavy amino acids during cell culture, then mix lysates |
Relative ratios in compatible cultured systems |
Chemical isotopic / isobaric labeling |
TMT, iTRAQ, dimethyl labeling |
Tag peptides after digestion; isobaric tags release reporters at MS2/MS3 |
Relative ratios inside a closed labeled set |
Older cysteine-directed tags |
ICAT-class reagents |
Enrich labeled cysteine peptides for pairwise comparison |
Relative; coverage limited by cysteine chemistry |
Label-free discovery |
DDA-LFQ, unlabeled DIA |
Independent injections; peak area or fragment chromatograms |
Relative matrix across a coherent run plan |
Targeted absolute |
AQUA/SIS with MRM or PRM |
Closed peptide list plus calibration |
Concentration in defined units |
Isobaric kit names that include “absolute” still report relative channel ratios unless matched standards and calibration are added.
Labeled Quantitative Methods
Metabolic labeling (SILAC)
SILAC grows cells in medium containing light or heavy essential amino acids so newly synthesized proteins become labeled. Lysates are mixed early, which can reduce some digestion variability for those channels. The method fits controlled cell-culture systems. It is usually impractical for many primary tissues, archival samples, and clinical biofluids.
Isobaric chemical labeling (TMT and iTRAQ)
TMT and iTRAQ label peptide N-termini and lysines with isobaric tags that share precursor mass and release distinct reporters after fragmentation (Thompson et al. 2003). Channels are pooled into one run, which supports same-run relative comparison inside a finite plex. Ratio compression from co-isolation remains a design limit and may need fractionation or MS3 strategies.
Dimethyl labeling and related chemical isotope tags
Dimethyl labeling marks amines with light or heavy dimethyl groups after digestion and is often used for cost-sensitive pairwise or small multiplex designs. Mixing occurs after digestion, so digestion and labeling completeness remain separate QC points. Reproducibility is not automatically equal to early-mix SILAC.
Historical or niche tags (ICAT, ¹⁸O)
ICAT-class reagents target cysteine-containing peptides and historically supported pairwise isotopic comparison with affinity enrichment. Coverage is bounded by cysteine content and enrichment chemistry; they should not be described as unrestricted whole-proteome absolute assays. Enzyme-catalyzed ¹⁸O labeling during digestion can support isotopic pairs in specialized designs, but exchange completeness and back-exchange need explicit control.
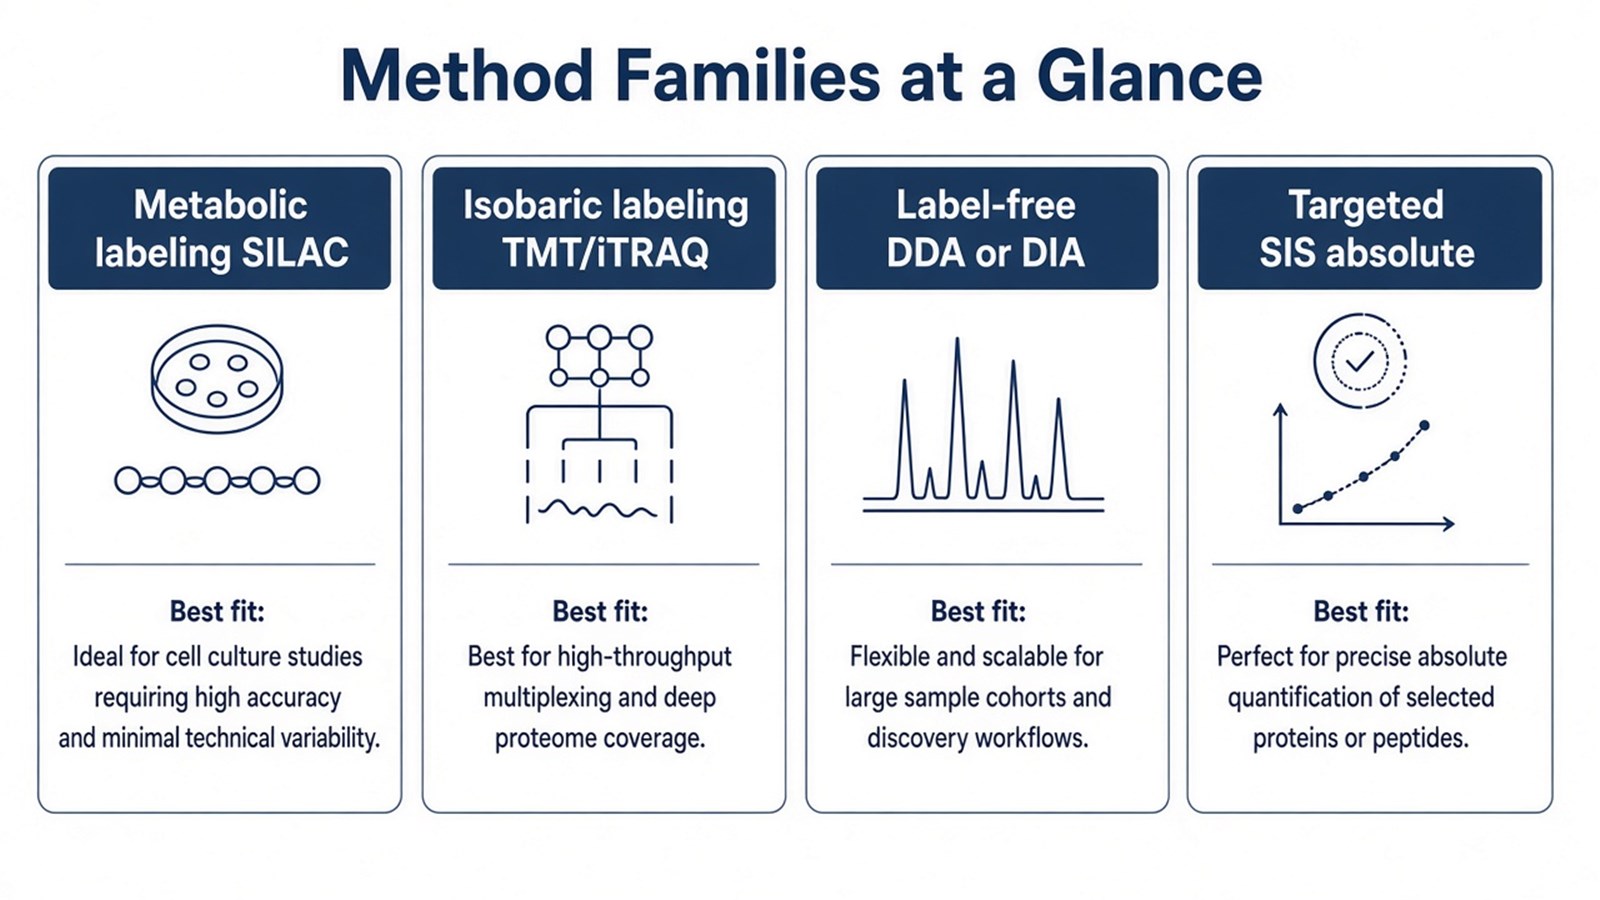
Figure 2. Common quantitative families differ by when labels enter the workflow and whether samples share one acquisition path.
Label-Free Quantitative Methods
LFQ omits isotopic or isobaric tags. Each sample is acquired independently by data-dependent acquisition (DDA) or data-independent acquisition (DIA).
Intensity-based LFQ (including MaxLFQ)
After peptide identification, software integrates precursor chromatograms, aligns runs, and normalizes to estimate relative protein change (Cox et al. 2014). This is the default discovery metric for many DDA label-free projects.
Spectral counting
Abundance is approximated from the number of assigned MS/MS spectra. It can support rough ranking but is usually weaker for subtle fold changes than peak-area LFQ.
Unlabeled DIA
DIA fragments scheduled precursor windows and recovers fragment chromatograms, including SWATH-style library routes (Gillet et al. 2012). DIA is an acquisition mode inside label-free chemistry, not a synonym for every LFQ project.
Judgment Criteria: Choosing Among the Methods
Study need |
Often prefer |
Reassess |
|---|---|---|
Cultured cells, early mixing useful |
SILAC |
Clinical tissue or biofluid that cannot be metabolically labeled |
Closed multi-condition panel, same-run comparison |
TMT or iTRAQ |
Open enrollment after mixing; absolute units required |
Independent unlabeled discovery, flexible N |
LFQ (DDA or DIA) |
Must freeze contrast in one plex |
Cost-sensitive small chemical multiplex |
Dimethyl or related tags |
Need high-plex commercial isobaric kits |
Concentration or specification units |
AQUA/SIS targeted assay |
Expect discovery TMT or LFQ tables to provide amounts |
Closed confirmation panel |
PRM or MRM |
Open proteome still required |
Prefer labeled multiplex when chemistry and plex membership fit. Prefer label-free when samples must stay independent. Prefer targeted SIS assays when the claim is absolute. Prefer discovery first, then targeted follow-up, when the protein list is still open.
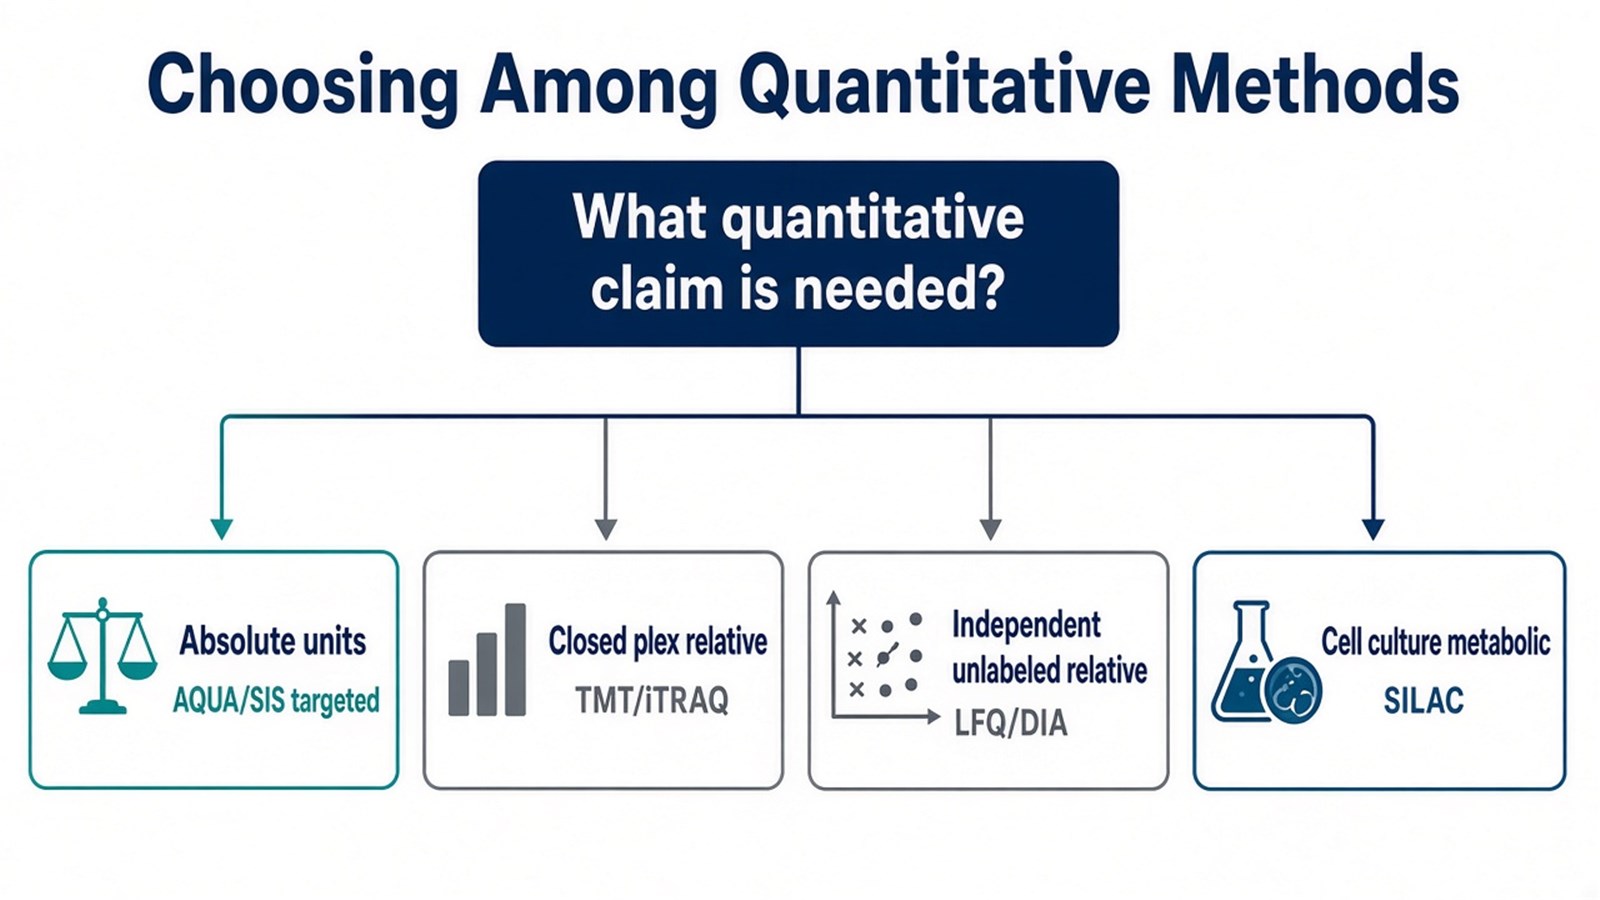
Figure 3. Start from the quantitative claim and sample constraints, then choose among labeled, label-free, or absolute targeted families.
For pairwise TMT-versus-LFQ selection detail, researchers can refer to TMT vs Label-Free Quantitative Proteomics and TMT vs Label Free. For absolute method families, see Absolute Protein Quantification: Methods and Uses.
Limits and Application Boundaries
No family removes the need for matched sample preparation and declared QC. Labeled methods add reagent cost and chemistry failure modes. Label-free methods add run-to-run drift and missing-value risk. Pathway enrichment on any discovery list reflects overrepresentation relative to a quantified background. It does not prove pathway activity. Diagnostic cutoffs sit outside ordinary research reporting unless a separately validated assay framework is in place.
Typical applications span cell-signaling screens (SILAC or LFQ), multi-condition drug panels (TMT/iTRAQ), clinical or archival discovery cohorts (LFQ/DIA), and biomarker concentration panels (SIS-targeted). Once sample type, enrollment rules, and the relative-versus-absolute claim are defined, the project can be mapped to a method family. Researchers can review the MtoZ Biolabs TMT Quantitative Proteomics Analysis Service, Label-Free Quantitative Proteomics Service, Deep DIA Proteomics Service, and Absolute Quantitative Analysis (AQUA) Service for feasibility and workflow planning.
Frequently Asked Questions
1. Are labeled methods always more accurate than label-free methods?
No. Accuracy depends on chemistry completeness, co-isolation or batch control, and the stated claim. Labels change how comparison noise enters the experiment.
2. Does iTRAQ or TMT provide absolute quantification by default?
No. Despite historical naming, ordinary reporter ratios are relative inside the plex unless SIS calibration is added.
3. Is DIA a label-free method or a separate third category?
DIA is an acquisition mode. It is commonly paired with label-free sample handling, but labeling and acquisition remain separate decisions.
4. When is SILAC preferred over TMT?
When metabolic labeling in culture is feasible and early lysate mixing helps the design. TMT is often preferred when tissues or biofluids must be chemically labeled after digestion.
5. How should absolute assays sit in this overview?
As a separate claim family for closed targets with standards, not as a drop-in replacement for discovery TMT or LFQ tables.
Reference
- A. Thompson, J. Schafer, K. Kuhn, S. Kienle, J. Schwarz, G. Schmidt, T. Neumann, C. Hamon (2003). Tandem mass tags: a novel quantification strategy for comparative analysis of complex protein mixtures by MS/MS. Anal. Chem., 75, 1895-1904. https://doi.org/10.1021/ac0262560
- J. Cox, M.Y. Hein, C.A. Luber, I. Paron, N. Nagaraj, M. Mann (2014). Accurate proteome-wide label-free quantification by delayed normalization and maximal peptide ratio extraction, termed MaxLFQ. Mol. Cell. Proteomics, 13, 2513-2526. https://doi.org/10.1074/mcp.M113.031591
- L.C. Gillet, P. Navarro, S. Tate, H. Rost, N. Selevsek, L. Reiter, R. Bonner, R. Aebersold (2012). Targeted data extraction of the MS/MS spectra generated by data-independent acquisition: a new concept for consistent and accurate proteome analysis. Mol. Cell. Proteomics, 11, O111.016717. https://doi.org/10.1074/mcp.O111.016717
