



Chemical cross-linking mass spectrometry quality control decides whether a reported link can be used as a distance restraint. The experiment covalently joins residues that are close enough and reactive enough. The search then proposes cross-linked peptide-spectrum matches. Acceptance is a later step. A gel shift is not a residue pair. A long hit list is not a structure. Each accepted link still needs a declared identification FDR, a site call or an ambiguity flag, and fragment support from both peptides. Buffer amines and lysine access belong in the same plan as the search FDR. Only then can the pair be written as a spacer-limited constraint.
Researchers who already have a named pair or complex, an amine-free buffer, and a written restraint claim can review the MtoZ Biolabs Crosslinking Mass Spectrometry Service for feasibility. Method-level choice among enrichment and residue-level routes is covered in Measuring Protein-Protein Interactions.
XL-MS Quality Control Accepts Links as Restraints
An XL-MS project asks two jobs. The reaction asks whether the chosen reagent, often DSS, BS3, or DSSO, formed covalent products in the submitted buffer. The detection job identifies linked peptides after digestion. Quality control sits between those jobs. It asks which proposed links survive efficiency, FDR, localization, and spectral review.
Do not treat a crosslink table as an atomic model. Leitner and colleagues describe XL-MS as an integrated source of residue-pair restraints for molecular machines, not as a replacement for a unique coordinate set (Leitner et al. 2016). An inter-protein link supports a contact hypothesis within reagent reach. An intra-protein link constrains fold or conformation. A missing link does not exclude contact. Lysines may be absent, buried, or lost after digestion.
If the next sentence must name a recovered pair in lysate, use a later Co-IP or AP-MS arm rather than stretch the restraint table. Enrichment follow-up is scoped through the IP-Mass Spectrometry Service. If the next sentence is a docked pose, keep the accepted links as restraints on the Protein-Protein Interface Analysis Server workflow, not as proof that one decoy is correct.
Cross-Linking Efficiency Before a Search Is Trusted
Cross-linking efficiency is a sample check. It asks whether enough productive links formed without collapsing the protein into an uninterpretable smear. SDS-PAGE can show residual monomer and a higher-mass product when an oligomer or complex is expected. A complete disappearance of monomer can mean over-reaction. An unchanged ladder can mean the reagent was quenched, the buffer contained amines, or accessible lysines were scarce.
Mass spectrometry adds a second efficiency view. Dead-end or mono-links show that the reagent reached the protein but found no partner residue. Intra-protein links show local proximity. Inter-protein links are the species later used as pair restraints. A search dominated by dead-ends is not a failed instrument run. It is a warning that the reaction or the interface lysine set did not produce pair-level products.
Tris or glycine must be exchanged before the reaction. Those amines consume NHS esters. Planning amounts for purified proteins are 100 to 150 ug at purity of at least 90 percent, with 20 ug used only when material is limited. Cell-level crosslinking typically plans 1x10^7 to 2x10^7 cells per condition. Solutions and cells travel snap-frozen on dry ice after quench.
If the interface is lysine-poor or the interaction is too short for a solution NHS reaction, chemical XL-MS efficiency will stay low. Photo-activated capture can then be reviewed through the Photocrosslinking Mass Spectrometry Service.
Cross-Linked Peptide FDR and Site Localization
A protein FDR used in shotgun proteomics is not the same filter. Cross-linked peptide-spectrum matches need their own target-decoy estimate because two peptides and a linker mass are scored together (Walzthoeni et al. 2012). Acceptance should state the FDR definition, the decoy scheme, and whether the cutoff was applied to unique residue pairs or to every spectrum match.
Passing FDR does not finish site localization. A peptide can contain more than one lysine. The search may report a pair of peptides while leaving the exact residues ambiguous. An accepted table should mark localized sites and ambiguous peptides as different rows. Ambiguous rows can still show that two chains were covalently joined. They should not be drawn as a single C-alpha to C-alpha restraint.
Search in this intake typically uses pLink 2 or MaxQuant, and the stronger crosslink result is delivered. Software choice does not replace the FDR statement. A count of all raw hits before decoy filtering is not an accepted result.
The comparison below keeps reaction checks and identification checks on separate tracks.
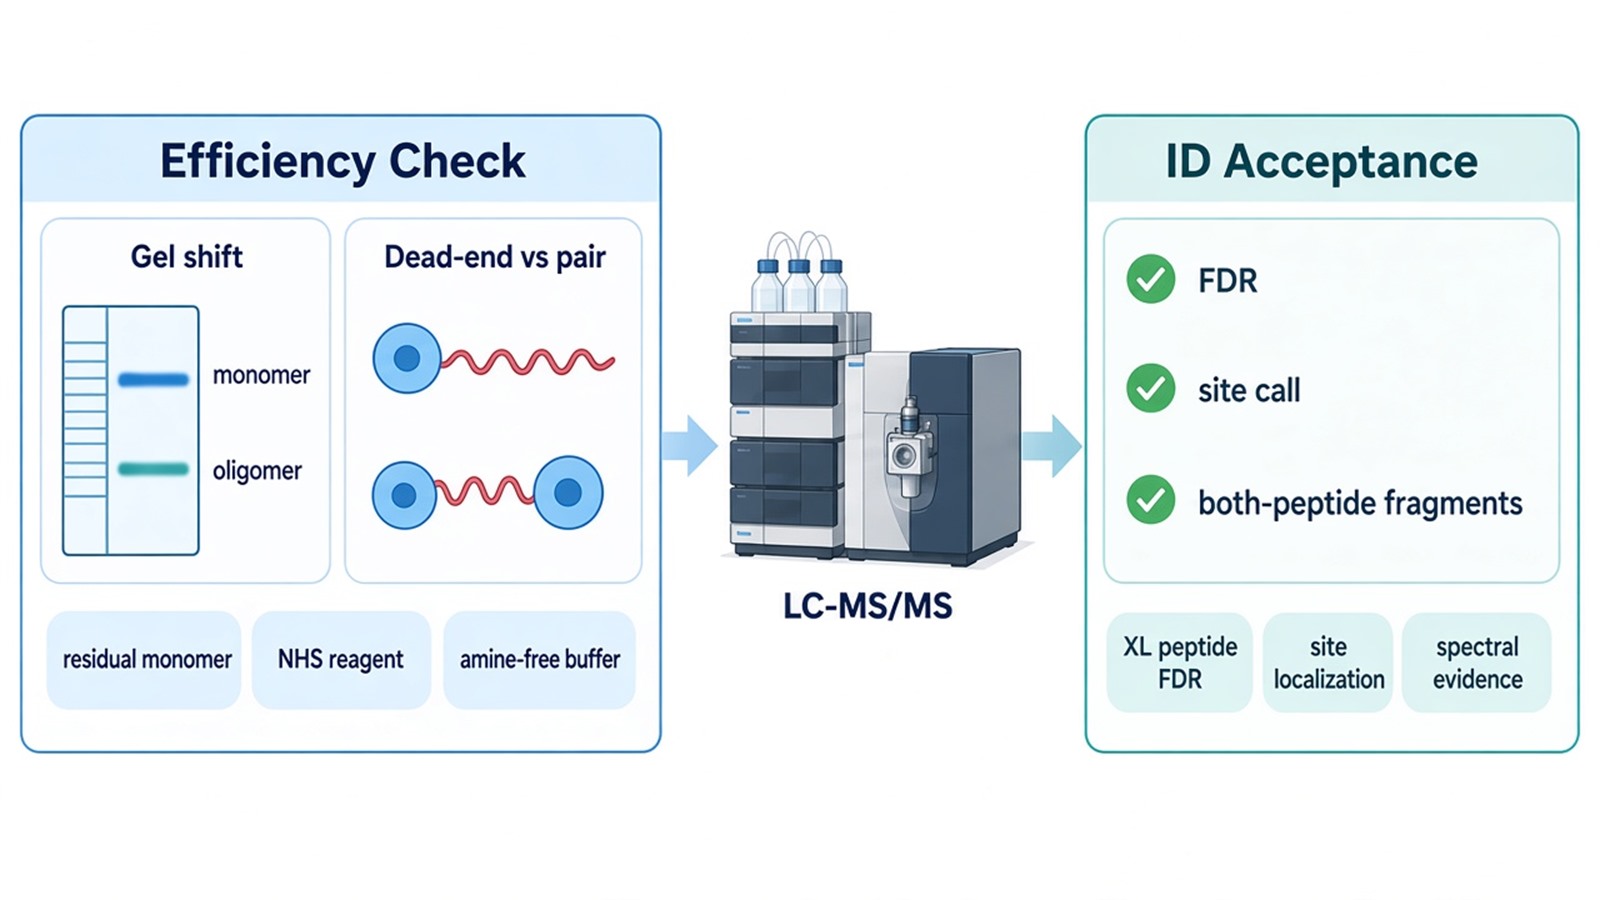
Figure 1. Cross-Linking Efficiency vs Identification Acceptance.
Spectral Evidence Required for Acceptance
Each accepted link needs fragment support from both peptides. A precursor mass that matches a linker-shifted pair is not enough. Sequence ions should assign residues on each arm. For a cleavable reagent such as DSSO, linker-specific fragment ions can add a second check that the two peptides were joined by that chemistry. For a non-cleavable reagent such as DSS or BS3, both sequences still need fragment coverage after the pair is scored.
Reject a row when only one peptide is explained, when the spectrum is dominated by a single abundant ion with no sequence ladder, or when the same spectrum is better explained as a mono-link or a linear peptide. Intra-protein and inter-protein classes must stay labeled. Mixing them inflates a pair-contact table.
Relative intensity can be compared across conditions when matched aliquots were labeled and searched the same way. Intensity does not convert a weak spectrum into a localized site.
Distance Constraints After QC, Samples, and Deliverables
An accepted inter-protein link is a distance constraint. The spacer of the chosen reagent plus the two side chains sets the reach. The constraint can support or reject a pose. It cannot write atomic coordinates. HDX-MS reports peptide-level protection, not the same residue-pair geometry. Protection follow-up is scoped through the [Protein-Protein Interactions Characterization Service |
HDX-MS](https://www.mtoz-biolabs.com/protein-and-protein-binding-site-analysis.html). |
|---|
XL-MS quality control fails a restraint when efficiency is unchecked, when FDR is undeclared, when the site is ambiguous but drawn as unique, or when the spectrum does not support both arms. It also fails when the publication sentence is a unique structure or a cellular affinity value.
QC item |
What is inspected |
Accepted use |
Not accepted as |
|---|---|---|---|
Cross-linking efficiency |
Gel shift and dead-end versus pair products |
Evidence that the reaction occurred |
A unique pose |
Cross-linked peptide FDR |
Target-decoy of XL matches |
Ranked residue-pair table |
Every raw search hit |
Site localization |
Which residues carry the linker |
Localized pair or flagged ambiguity |
A complete interface map |
Spectral evidence |
Fragments from both peptides |
One accepted link |
A protein ID without the link |
Distance constraint |
Spacer reach versus a model |
A restraint on architecture |
Atomic coordinates |
The table is an acceptance rule, not a ranking of reagents.
Ship already crosslinked gel bands, protein solutions, or cells, or ship two uncrosslinked recombinant proteins with sequences. State the reagent, quench, buffer amines, and whether a model will be used only to interpret links. Orbitrap-class high-resolution LC-MS/MS is the usual acquisition platform. Database search should use the client sequences or UniProt.
Intake is faster when the QC items above are named before shipment. The diagram below links buffer, amount, efficiency, FDR, spectra, and the restraint claim to an XL-MS request.
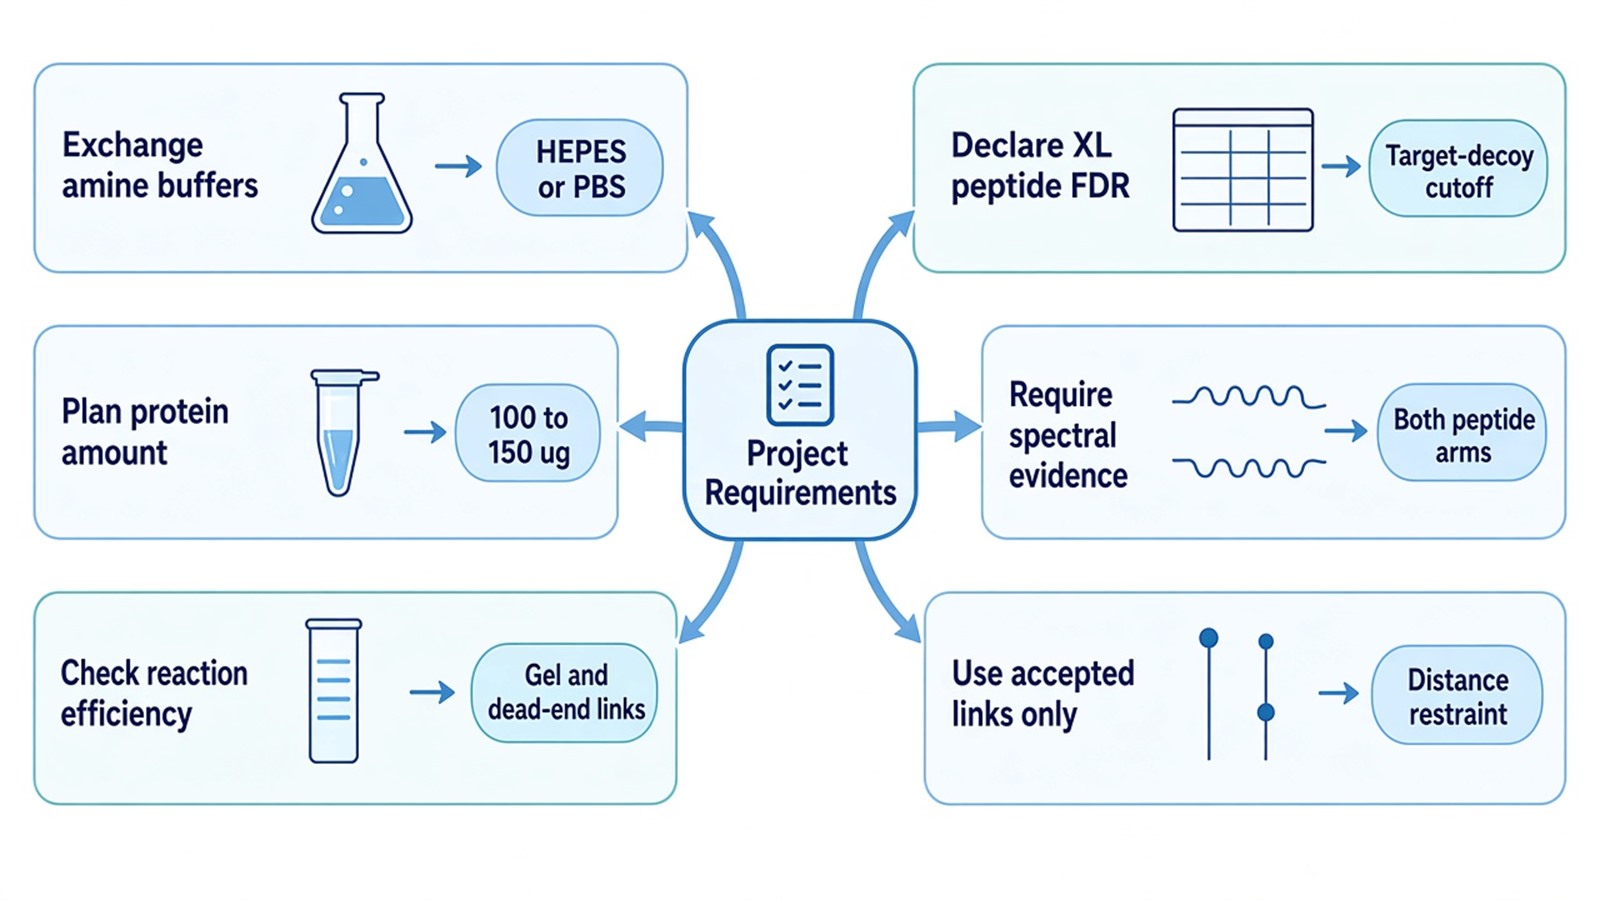
Figure 2. Matching Project Requirements With XL-MS Quality Control.
Deliverables typically include a residue-pair table split into intra-protein and inter-protein classes, FDR and localization flags, annotated spectra for accepted links, and a short methods note on reagent and search settings. Deliverables do not include a unique atomic model, a guaranteed inter-protein count, or a claim of cellular binding energy. To start intake, send the pair or complex, sequences, buffer, planned reagent, and intended restraint sentence to info@MtoZ-Biolabs.com or use the site inquiry form so the QC path can be confirmed before shipment.
Frequently Asked Questions
1. Is an SDS-PAGE shift enough to accept the residue-pair table?
No. A shift shows that a covalent product formed. Acceptance of each row still requires a declared XL FDR, a site call or ambiguity flag, and fragment support from both peptides.
2. Does passing FDR mean every residue pair is uniquely localized?
No. FDR ranks matches. Localization assigns the linked residues. A peptide with two lysines can pass FDR and still be ambiguous.
3. Can protein stored in Tris or glycine be submitted for NHS crosslinking?
Not without buffer exchange. Those amines consume DSS, BS3, or DSSO. Plan HEPES or PBS and declare any leftover amine.
4. How much purified protein is typically planned for chemical XL-MS?
Plan 100 to 150 ug at purity of at least 90 percent. 20 ug is a limited-material fallback, not the first plan. Cell crosslinking typically plans 1x10^7 to 2x10^7 cells per condition.
5. If no inter-protein links pass QC, should the project move to photo-crosslinking?
Often yes when lysines are scarce or the pair is too short-lived for a solution NHS reaction. First confirm that buffer amines and over- or under-reaction were not the cause.
Researchers can separate reaction checks from identification acceptance, then confirm reagent, sequences, and QC flags through the MtoZ Biolabs Crosslinking Mass Spectrometry Service.
Reference
- A. Leitner, M. Faini, F. Stengel, R. Aebersold (2016). Crosslinking and mass spectrometry: an integrated technology to understand the structure and function of molecular machines. Trends Biochem. Sci., 41, 20-32. https://doi.org/10.1016/j.tibs.2015.10.008
- T. Walzthoeni, M. Claassen, A. Leitner, F. Herzog, S. Bohn, F. Forster, M. Beck, R. Aebersold (2012). False discovery rate estimation for cross-linked peptides identified by mass spectrometry. Nat. Methods, 9, 901-903. https://doi.org/10.1038/nmeth.2103
