



Cellular IP product mass spectrometry detection starts after an immunoprecipitation has already been run, not after a textbook definition of IP or MS. The submitted material is an eluate, washed beads, or a stained gel band from a cell IP or Co-IP. LC-MS/MS then identifies proteins that survived capture and washing. The report is an identification table compared with negative controls. It is not a structure, not a predicted interactome, and not a global expression profile of the starting lysate. Digestion path and contaminant filters should be chosen from the buffer and control set that actually arrive with the tubes.
Send the IP history with the tubes. Antibody catalog information, bead type, lysis detergent, wash salt, elution chemistry, and which control lanes were processed in parallel decide whether digestion can proceed. Teams with bait IP, matched IgG or empty-bead controls, and a written claim can review MtoZ Biolabs IP-Mass Spectrometry Service for feasibility. Pair-level method choice after a candidate list is covered in Measuring Protein-Protein Interactions.
What Must Be Declared With a Cellular IP Product
For cellular IP product mass spectrometry detection, the product is defined by the capture, not by the cell line name alone. State whether the bait was recovered with a native antibody or a tag, whether Protein A/G magnetic beads or agarose were used, and whether the tube contains eluted protein or still-bound beads. State the elution buffer if the complex already left the resin. SDS, high salt, glycine, urea, and residual NP-40 or Triton change the digestion path.
Declare the control set at the same time. An isotype IgG IP, a bead-only capture, or an empty-vector tag IP processed through the same wash and elution is the comparison that later filters the identification table. A single untreated lysate lane does not replace those IP-matched controls. If only the bait IP will be shipped, the report can list identified proteins in that tube. It cannot rank specific co-enriched partners.
Also declare whether the IP was run under native or denaturing lysis. Native Co-IP can keep partners that washing still allows. Denaturing IP is closer to bait recovery plus sticky background. Cellular IP product mass spectrometry detection cannot reconstruct the lysis condition from a peptide list.
Eluate Digestion Versus On-Bead Digestion
Two handoff formats cover most cellular IP products. In-solution digestion fits a liquid eluate that can be reduced, alkylated, and cut with trypsin after interfering buffer is removed. On-bead digestion fits washed magnetic beads when elution would add SDS or cause transfer loss (Turriziani et al. 2014). In-gel digestion fits a stained band cut from SDS-PAGE of an IP eluate.
Choose the path from the tube that will arrive, not from a preferred instrument name. Acid eluates and volatile buffers are easier to take into in-solution digest after a short cleanup. SDS sample buffer and high-salt eluates need dilution, precipitation, or trap-based cleanup before trypsin. Beads should arrive washed and without large volumes of free antibody storage buffer. Gel bands should use Coomassie, SYPRO Ruby, or an MS-compatible silver stain; destaining history should be written on the form.
On-bead digestion does not remove the need for controls. Antibody and bead proteins still enter the peptide pool. The advantage is that the complex never has to leave the resin in a detergent slug. The limit is that residual detergent or primary amine buffers on the beads can still suppress digestion or later ionization.
Eluate and on-bead routes meet at the same peptide cleanup step. The comparison below groups those handoffs before desalting and LC-MS/MS.
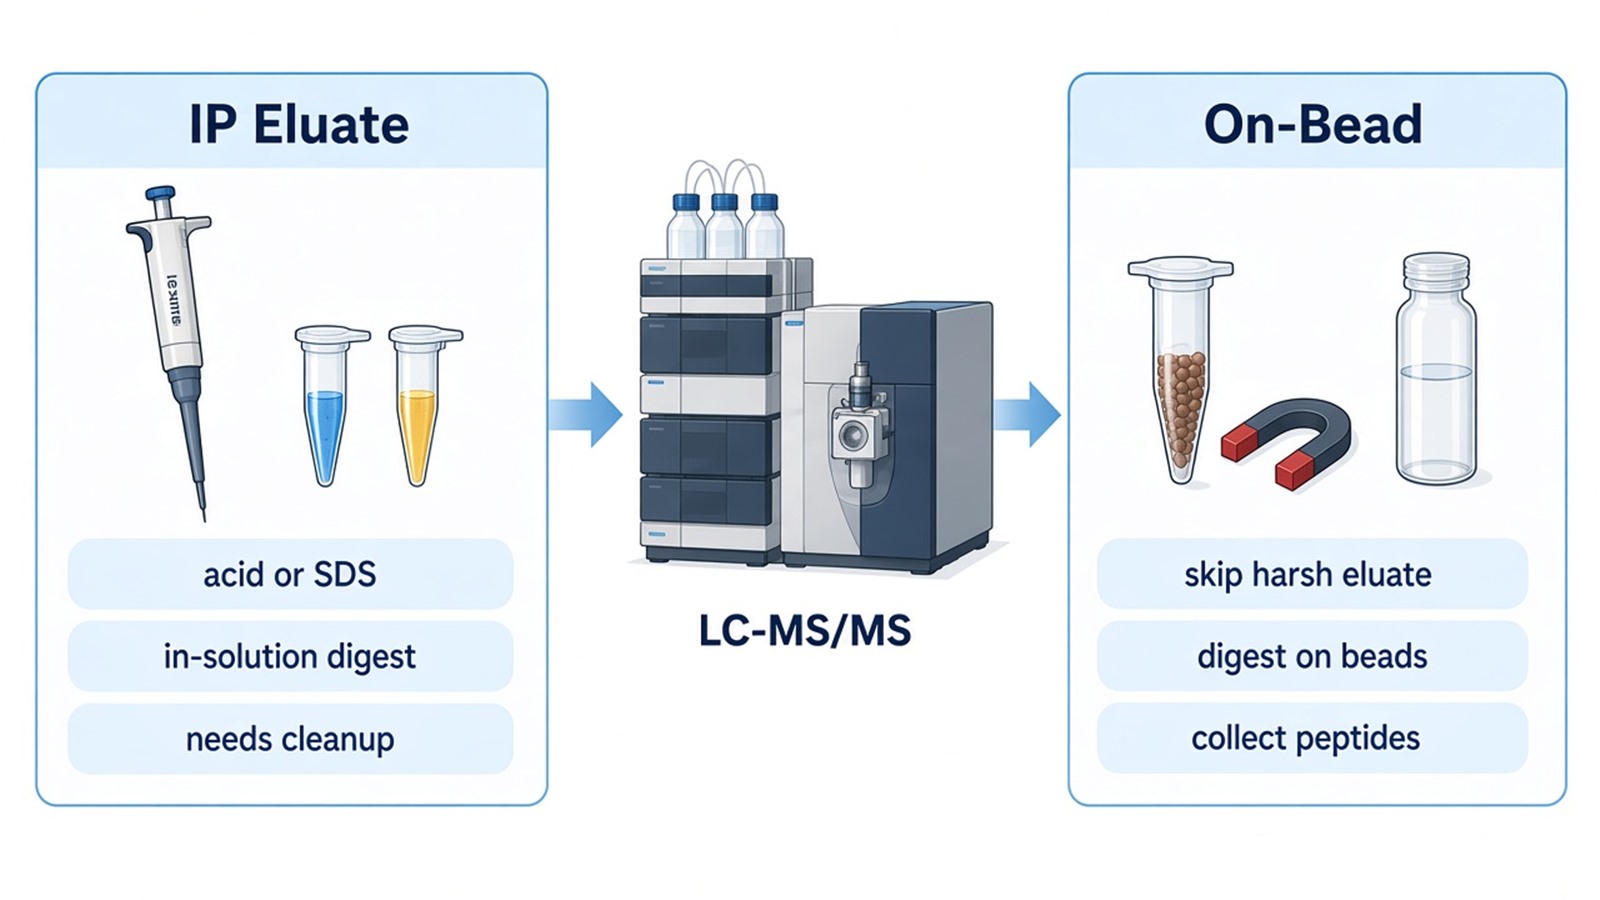
Figure 1. IP Eluate Digest vs On-Bead Digest.
Cleanup, Desalting, and LC-MS/MS
Peptides from a cellular IP product usually need desalting or a trap clean-up before nanoLC. Residual SDS, NP-40, high NaCl, and excess glycine suppress ionization and foul the column. Stage-tip, spin-column, or S-Trap style cleanup is chosen from the buffer that was declared, not as a hidden extra step after a dirty eluate is accepted without comment.
Liquid IP products above roughly 10 ug total protein are easier to process than low-microgram tubes that still contain SDS or salt. Below that range, detergent and salt should be reduced before shipment, or the project should switch to on-bead or gel-band format. Buffer composition and an estimated protein amount should be written on the intake form. Compatible gel-band rules for pull-down and IP eluates are the same as those listed for the Fusion Protein Interaction Analysis Service | Pull-Down and MS.
LC-MS/MS then records peptide m/z and fragment spectra for database search. The search identifies proteins present in that digest. It does not, by itself, mark them as interactors. Acquisition depth should match a typical IP eluate, which is simpler than a whole-cell proteome and still contains antibody chains, keratins, and abundant background. PTM site tables are a separate scope and should be requested only when the IP and the digest were planned for that modification.
Negative Controls and Contaminant Filtering
Negative controls are part of cellular IP product mass spectrometry detection, not an optional bioinformatics add-on. The control IP should use the same cell input, bead lot, wash, elution or on-bead digest, and LC-MS/MS method. IgG or empty-bead controls address antibody and resin binding. Empty-tag or bait-depleted inputs address expression artifacts when a tagged bait was used.
Filtering has two layers. Experimental controls remove names that are equally present in the negative IP. Catalogs such as the CRAPome flag proteins that recur across many affinity purifications, which helps interpret leftovers after control subtraction (Mellacheruvu et al. 2013). Scoring tools such as SAINT can rank bait IPs against a set of controls when replicates exist (Choi et al. 2011). A CRAPome label without a matched control IP is not a substitute for that control.
Keratin, trypsin, and antibody sequences are expected. They should be annotated, not presented as novel partners. Ribosomal proteins, heat-shock proteins, and highly abundant metabolic enzymes often survive IP washes. They remain on the table until control enrichment says otherwise. End-to-end capture-to-search logic for Co-IP-MS is summarized in How to Combine Co-IP with Mass Spectrometry for Protein Interaction Analysis.
Identification Tables and What They Can Support
The primary deliverable is an identification table per submitted IP and control: protein accession, gene name, peptide counts or intensity, and a note if the protein is an expected contaminant. When controls were searched, a second table can list proteins enriched with the bait versus those controls. Presence in the bait IP alone is not a sufficient call.
|
Table column or filter |
What it supports |
What it does not support |
|---|---|---|
|
Protein ID in the bait IP |
The protein was in that digest |
Specific interaction with the bait |
|
Peptide count or intensity vs control IP |
Enrichment under the submitted wash |
Direct binding or stoichiometry |
|
CRAPome or recurrent-contaminant flag |
Common affinity-purification background |
Automatic removal without a control |
|
Antibody or keratin entries |
Expected reagent and handling proteins |
A failed IP |
|
Optional PTM site table |
Modification evidence when scoped |
A default output of every IP-MS run |
The table is a candidate list for follow-up, not a finished interaction map. Co-enrichment can still be indirect (Hein et al. 2015). Reciprocal IP, a defined-pair pull-down, or an orthogonal assay is required if the next sentence will claim a named pair. Mixed enrichment-MS projects that include client-prepared IP products can also be reviewed under the MS-Based Protein-Protein Interaction Analysis Service.
Do not treat a long identification list as better detection. After cleanup and control filtering, a shorter enriched set is the usable result. If the bait itself is missing from the bait IP table, stop and review antibody, elution, and digest before interpreting partners.
Samples, Deliverables, and Feasibility Intake
Ship materials for cellular IP product mass spectrometry detection with a short form: species and cell type, bait and antibody, bead type, lysis and wash buffer, elution or on-bead status, estimated protein amount, stain type for gel bands, and which tubes are negative controls. Keep beads cold and without drying out. Avoid extra freeze-thaw of dilute SDS eluates. If elution chemistry is still open, buffer options that are more compatible with later cleanup are discussed in What Elution Buffer Should Be Used for Beads in Co-IP Samples Before Mass Spectrometry.
Intake is faster when the product format is mapped before shipment. The diagram below links common IP-product states to digest path and control filtering.
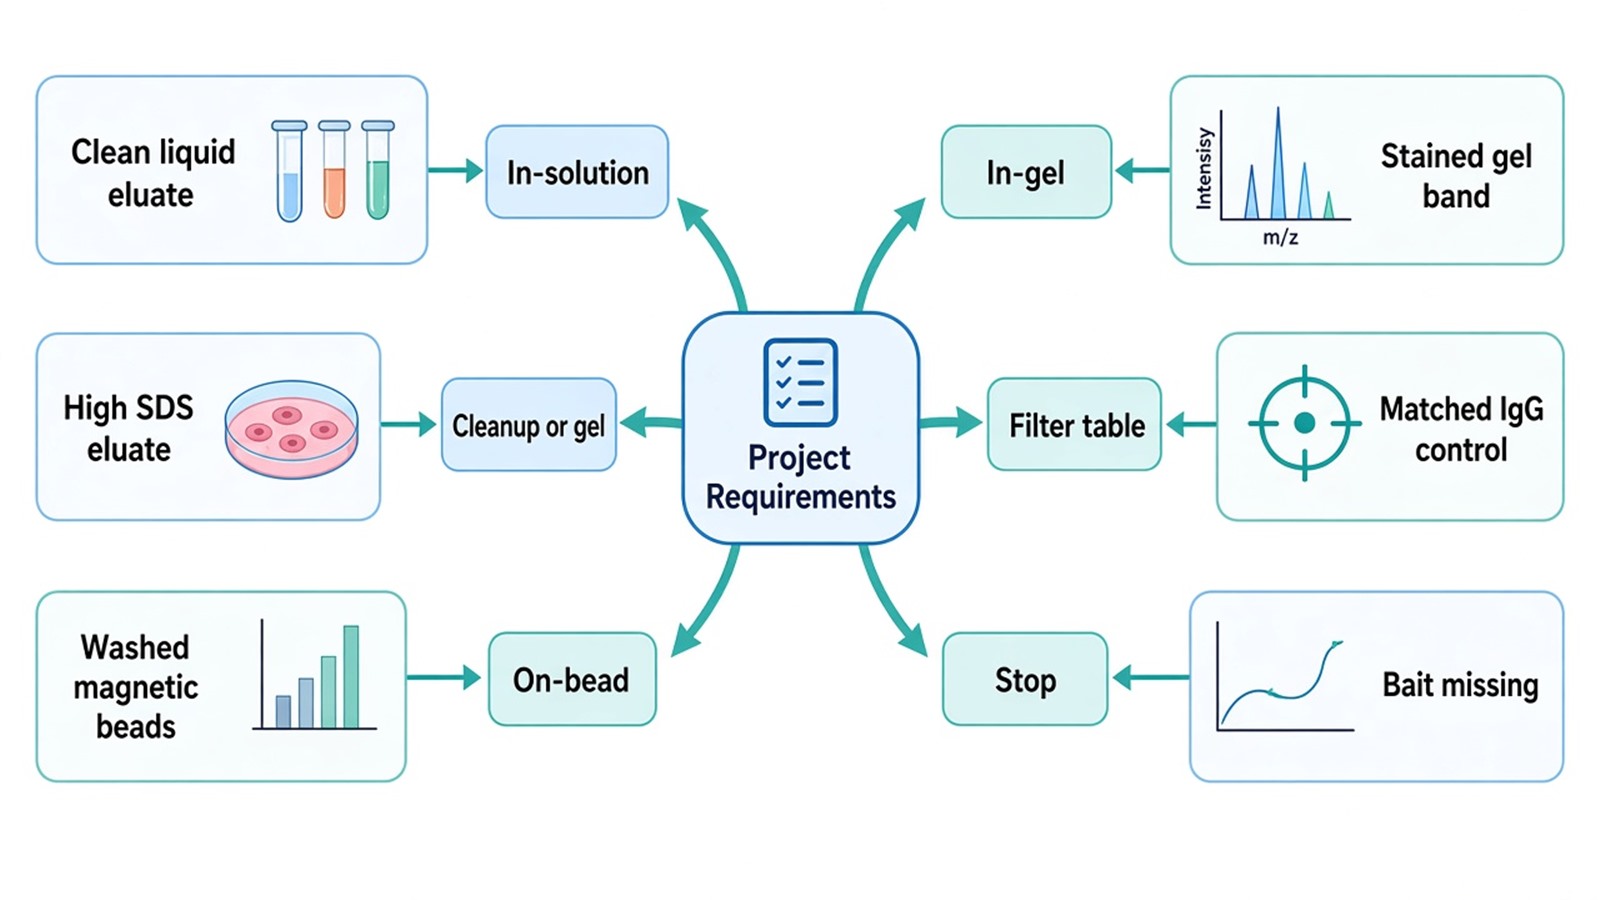
Figure 2. Matching Cellular IP Products With LC-MS/MS Intake.
Deliverables typically include processed identification tables, a control-aware candidate list when negatives were submitted, and a short methods note on digest and search settings. Raw files can be discussed when a journal archive is required. Deliverables do not automatically include a network figure as proof, a PTM atlas, or a binding-site map. To start intake, send the bait, IP format, control plan, and buffer list to info@MtoZ-Biolabs.com or use the site inquiry form so the digest path can be confirmed before shipment.
Frequently Asked Questions
1. Can an SDS elution from Protein A/G beads be used for LC-MS/MS?
It can be considered when the SDS load can be reduced by precipitation, trapping, or a compatible cleanup, or when a gel band is cut instead. High SDS in a low-microgram liquid tube can block digestion. Declare the SDS percentage and protein estimate before shipping.
2. Should washed beads or an eluate be sent for cellular IP product mass spectrometry detection?
Send washed beads when elution would add detergent or when protein amount is low and transfer loss is a concern. Send a liquid eluate when the buffer is already MS-friendly and protein is above the low-microgram, high-detergent range. Do not send both formats of the same IP unless they are labeled as technical splits.
3. Is an IgG control required if the CRAPome will be used?
Yes, if the claim is enrichment versus background in this experiment. CRAPome flags recurrent names across many studies. It does not measure antibody or bead binding in the current wash protocol.
4. How much IP product is typically needed?
Liquid samples above roughly 10 ug total protein are easier to digest and desalt. Lower amounts can be attempted after detergent and salt are reduced, or after switching to on-bead or gel-band input. Exact feasibility still depends on bait abundance and buffer.
5. What should the feasibility email list for a cell IP already sitting on beads?
List bait, antibody, bead type, last wash buffer, whether a matched IgG or bead-only tube exists, and whether on-bead digestion is acceptable. Feasibility review will not invent a control that was not performed.
Researchers submitting a completed cell IP can align eluate, bead, or gel-band format with control tubes, then confirm the digest path through MtoZ Biolabs IP-Mass Spectrometry Service intake.
Reference
- B. Turriziani, A. Garcia-Munoz, R. Pilkington, C. Raso, W. Kolch, A. von Kriegsheim (2014). On-beads digestion in conjunction with data-dependent mass spectrometry: a shortcut to quantitative and dynamic interaction proteomics. Biology, 3, 320-332. https://doi.org/10.3390/biology3020320
- H. Choi, B. Larsen, Z.Y. Lin, A. Breitkreutz, D. Mellacheruvu, D. Fermin, Z.S. Qin, M. Tyers, A.C. Gingras, A.I. Nesvizhskii (2011). SAINT: probabilistic scoring of affinity purification-mass spectrometry data. Nat. Methods, 8, 70-73. https://doi.org/10.1038/nmeth.1541
- D. Mellacheruvu, Z. Wright, A.L. Couzens, J.P. Lambert, N.A. St-Denis, T. Li, Y.V. Miteva, S. Hauri, M.E. Sardiu, T.Y. Low, et al. (2013). The CRAPome: a contaminant repository for affinity purification-mass spectrometry data. Nat. Methods, 10, 730-736. https://doi.org/10.1038/nmeth.2557
- M.Y. Hein, N.C. Hubner, I. Poser, J. Cox, N. Nagaraj, Y. Toyoda, I.A. Gak, I. Weisswange, J. Mansfeld, F. Buchholz, et al. (2015). A human interactome in three quantitative dimensions organized by stoichiometries and abundances. Cell, 163, 712-723. https://doi.org/10.1016/j.cell.2015.09.053
