



A laboratory applies tandem mass tag (TMT) labeling when several digests must be compared as relative protein ratios inside one liquid chromatography-tandem mass spectrometry (LC-MS/MS) injection. Peptides from each sample receive a channel-specific isobaric tag, the channels are mixed, and reporter ions released after fragmentation carry the channel abundances (Thompson et al. 2003). The useful application question is not which disease field mentions TMT. It is which study design can freeze a sample set at labeling and still answer a predefined group contrast.
TMT application is relative by default. It does not report mass or molar concentration unless a matched stable isotope-labeled internal standard (SIS) is added for that peptide. Labeling chemistry and acquisition mode remain separate decisions. In MtoZ Biolabs offerings, TMT projects are typically acquired as data-dependent acquisition (DDA) so reporters can be read at MS2 or MS3.
Study Designs Where TMT Labeling Is Commonly Applied
TMT labeling is commonly applied to treatment-versus-control sets, multi-condition panels, and time-course series when every planned harvest can be labeled into one plex, or into linked plexes that share a bridge or reference channel. Matched phenotype groups with similar matrix type also fit when biological replicates and channel layout are declared before tagging.
Those designs share one requirement. The comparison set is known early enough that amines can be labeled, quenched, and mixed before acquisition. Shared chromatography then removes run-to-run retention-time drift for that mixed set. The same requirement excludes designs that keep adding samples after mixing, or that only ask whether an unknown protein is present.
TMT labeling is applied to relative contrasts on a freezable sample set; kit channel count is not a biological result.
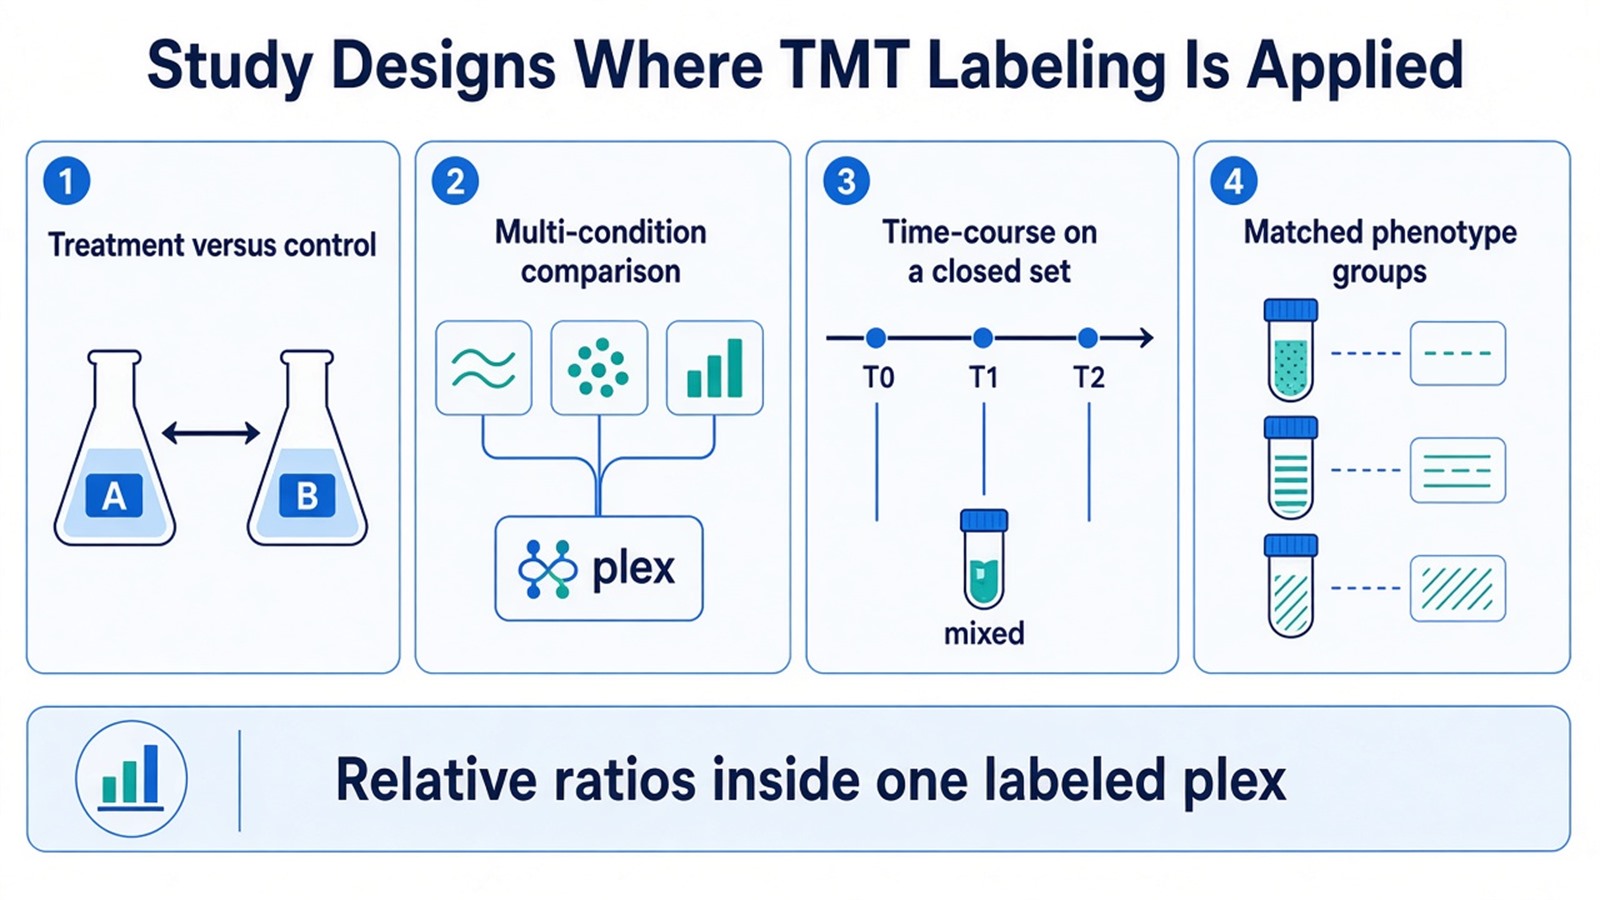
Figure 1. TMT labeling is applied when group structure can be mapped onto channels before peptides are mixed.
Discovery protein lists can still be open inside a TMT plex. What must be closed is membership of the plex. Researchers can refer to What Is TMT Isobaric Labeling? for the isobaric precursor and reporter-ion definition behind such applications.
How an Application Moves from Digests to a Relative Matrix
An applied TMT project follows a fixed order even when the biological theme changes. Proteins are extracted under matched conditions, digested, and checked for comparable load. Each digest receives one TMT channel. After quenching and cleanup, labeled peptides are pooled. Optional fractionation can deepen identification. LC-MS/MS then identifies peptides and records reporters. Analysis summarizes channel ratios to proteins and tests the predefined contrast.
Application success depends on that order more than on the disease keyword attached to the project. Unequal input, residual amine buffer, or incomplete labeling creates channel offsets that look like treatment effects after mixing. Co-isolated precursors at MS2 can compress true fold changes (Ting et al. 2011). An additional MS3 reporter scan can reduce that mixing on compatible instruments. It does not repair unmatched loads.
Reporter extraction and protein rollup are analysis steps. Researchers can refer to Principle of TMT-Based Quantitative Proteomics Analysis when the remaining question is how identification couples to reporter ratios.
Parameters That Decide Whether the Application Holds
Channel map and contrast must be written before labeling. Which biological groups occupy which tags, where replicates sit, and whether a bridge channel links plexes are design parameters. Leaving those decisions until after acquisition turns software into guesswork.
Matched protein input and digestion completeness decide whether reporter differences can be read as biology. Bulk assays remain useful for load matching. They are not the TMT result. Amine-free or low-amine buffers decide whether labeling chemistry can complete. Detergent and salt cleanup decide whether chromatography and spray stay stable for the mixed plex.
Acquisition parameters that matter for application include isolation purity for reporter scans and whether MS2 or MS3 is used for quantification. Analysis parameters include impurity correction, unique-peptide rules, within-plex normalization assumptions, and the exact statistical contrast. Pathway enrichment is optional after the ratio matrix passes quality checks. It is not a substitute for a failed plex design.
When extra samples are expected after the first harvest, unlabeled injections more often keep enrollment open. Researchers can refer to A Guide to Label-Free Quantitative Proteomics: From Experiment to Data Analysis for relative matrices built from separate runs.
Limits of Applying TMT Labeling in Those Study Contexts
TMT labeling does not inventory presence or absence of proteins that were never identified. A presence survey is a different claim from a same-protein ratio inside a plex. Ratio compression can shrink a real treatment effect when co-isolation is severe (Ting et al. 2011). A small observed fold change still needs isolation QC before it is called weak biology.
Application language such as biomarker discovery, mechanism study, or treatment response only holds when the assay returns a defensible relative matrix for predefined groups. It does not hold as a diagnostic claim, a therapy recommendation, or an absolute concentration. Multi-omics integration can use TMT ratios as one protein layer. It does not make TMT ratios interchangeable with transcript or metabolite units.
A later unlabeled digest cannot join the same reporter matrix. A new plex needs a bridge design. Targeted confirmation of a short named list is a follow-up application class, not an extension of TMT reporters.
How to Choose Whether TMT Labeling Fits the Next Project
The useful test is whether groups can freeze, whether digests can be amine-labeled cleanly, and whether the claim is a relative ratio.
Project need |
TMT labeling often fits |
Recheck or choose another route |
|---|---|---|
Treatment versus control on a fixed set |
Relative plex with matched load |
Samples still arriving after mixing |
Several conditions or time points |
Channels mapped before tagging |
Time points added after the plex is mixed |
Same matrix type across groups |
Shared chromatography after pooling |
Strongly unmatched matrices without a bridge plan |
Open enrollment cohort |
Unlabeled DDA or DIA injections |
Force late samples into an existing TMT mix |
Protein presence survey only |
Discovery identification plan |
Treat missing reporters as absence proof |
Named peptides need confirmation |
Targeted PRM or MRM follow-up |
Re-run the same TMT table as confirmation |
Amount or concentration unit |
SIS-supported targeted assay |
Ordinary TMT ratios read as ng per mL |
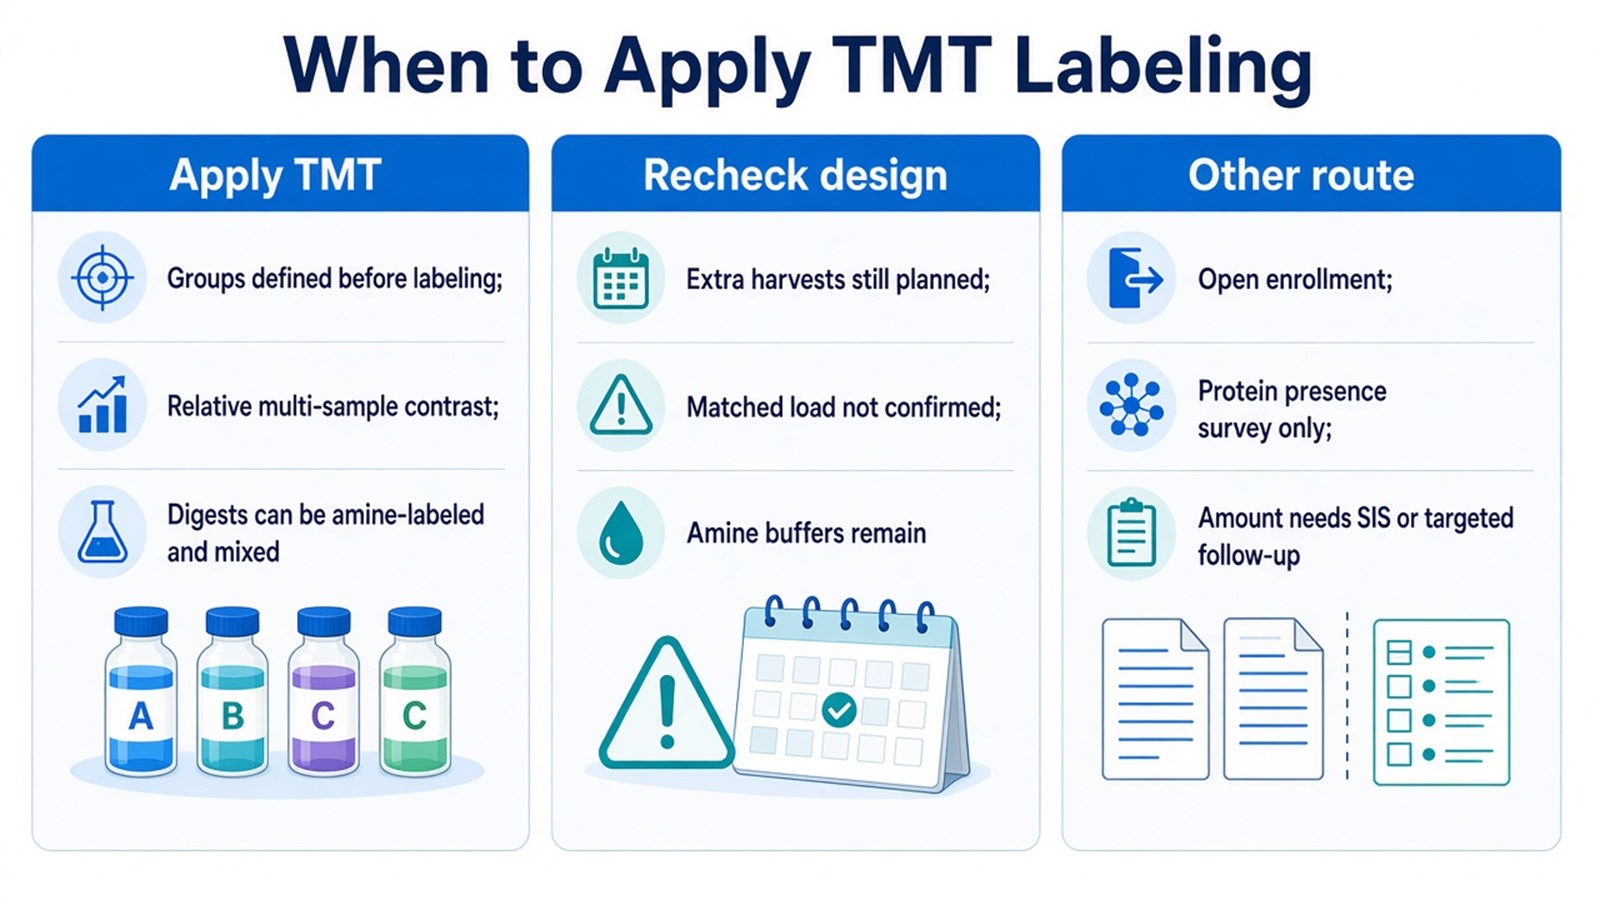
Figure 2. Apply TMT labeling when the sample set and relative contrast can be frozen before mixing; open enrollment and amount claims need another plan.
TMT more often fits a multi-group or closed time-course design that can be labeled together. An exception is a small, well-buffered set that still needs unlabeled flexibility because a follow-up harvest is already booked. Unlabeled label-free quantification more often fits open enrollment. An exception is a short closed cohort that must share one injection to limit run-order drift.
Researchers who already have defined groups, matched protein material, and a requested relative contrast can also review the MtoZ Biolabs TMT Quantitative Proteomics Analysis Service for project-specific plex planning.
Frequently Asked Questions
1. Is TMT labeling mainly an application for disease diagnosis?
No. TMT labeling supports relative protein ratios inside a mixed plex. Diagnostic use would require a separately validated clinical assay. A research contrast between disease-model and control samples is not a diagnostic claim.
2. Can TMT labeling replace a protein presence or absence screen?
No. Identification depth and search settings decide which proteins are named. Missing reporters do not prove that a protein is absent from the sample.
3. Does applying TMT always give fewer missing values than label-free quantification?
Mixing channels onto one chromatographic path can reduce run-to-run missingness for proteins that are identified in the mixed plex. It does not remove proteins that never yield an identification, and it does not repair incomplete labeling.
4. How should a treatment-response study use TMT if more doses will be added later?
Map the doses that exist now onto one plex, or plan a bridge sample for a second plex. Do not treat a later unlabeled digest as an extra TMT channel. If enrollment will stay open, unlabeled acquisition is often simpler.
5. Can TMT application results be used directly as absolute biomarker concentrations?
No. Reporter ratios are relative. Concentration reporting needs a matched SIS and a declared calibration, usually in a targeted follow-up assay.
MtoZ Biolabs can provide TMT labeling applications for predefined multi-group and time-course proteomics projects, including plex layout, labeling, LC-MS/MS, and relative matrix reporting when sample type and study design are available.
If further evaluation of channel mapping, bridge design, or whether unlabeled quantification fits better is needed, project details can be submitted for a feasibility review.
Reference
- A. Thompson, J. Schafer, K. Kuhn, S. Kienle, J. Schwarz, G. Schmidt, T. Neumann, R. Johnstone, A.K. Mohammed, C. Hamon (2003). Tandem mass tags: a novel quantification strategy for comparative analysis of complex protein mixtures by MS/MS. Anal. Chem., 75, 1895-1904. https://doi.org/10.1021/ac0262560
- L. Ting, R. Rad, S.P. Gygi, W. Haas (2011). MS3 eliminates ratio distortion in isobaric multiplexed quantitative proteomics. Nat. Methods, 8, 937-940. https://doi.org/10.1038/nmeth.1714
