



APEX proteomics is liquid chromatography–tandem mass spectrometry (LC-MS/MS) of proteins biotinylated by an engineered ascorbate peroxidase, usually APEX2, during a hydrogen peroxide (H₂O₂) pulse and then recovered on streptavidin. The file is a spatial or neighborhood proteome. It is not an accurate map of protein distribution, dynamics, and interaction networks under physiological conditions. High catalytic efficiency of APEX2 makes a short pulse possible. It does not confer superior sensitivity, lower background, or more precise localization than every other labeling method. An APEX-only arm, matched for localization, is part of the proteomic design. Without it, the streptavidin table is an uninterpreted biotinylated mixture.
Fusing APEX2 to a protein “involved in a drug mechanism” does not identify a drug target or validate a therapeutic partner. Precision-medicine and drug-discovery-pipeline language sits outside this assay. Chemical proteomics, in the usual activity-based-probe sense, is a different experiment. For the peroxidase chemistry that produces the biotin handle, APEX Proximity Labeling provides a complementary APEX2 discussion.
Researchers who already have an APEX2 fusion, a compartment tag, or a streptavidin eluate to search can also review the MtoZ Biolabs Proximity Labeling Mass Spectrometry Service for project-specific feasibility and analysis planning.
What the Proteome Contains
APEX2 converts biotin-phenol, not an epoxide, into a short-lived radical that marks nearby tyrosines. After the pulse, cells are lysed and biotinylated proteins are enriched. LC-MS/MS names those proteins and, if a quantitative method is used, compares them with APEX-only or a second pulse. Native structure and function are not preserved for the readout. The analyte is a digest of a streptavidin enrichment.
A whole-cell shotgun proteome after lysis reports abundance in the extract. An APEX proteome reports who was near the enzyme when H₂O₂ was added. Those are different tables. Calling the second table in-situ interaction profiling collapses proximity into binding.
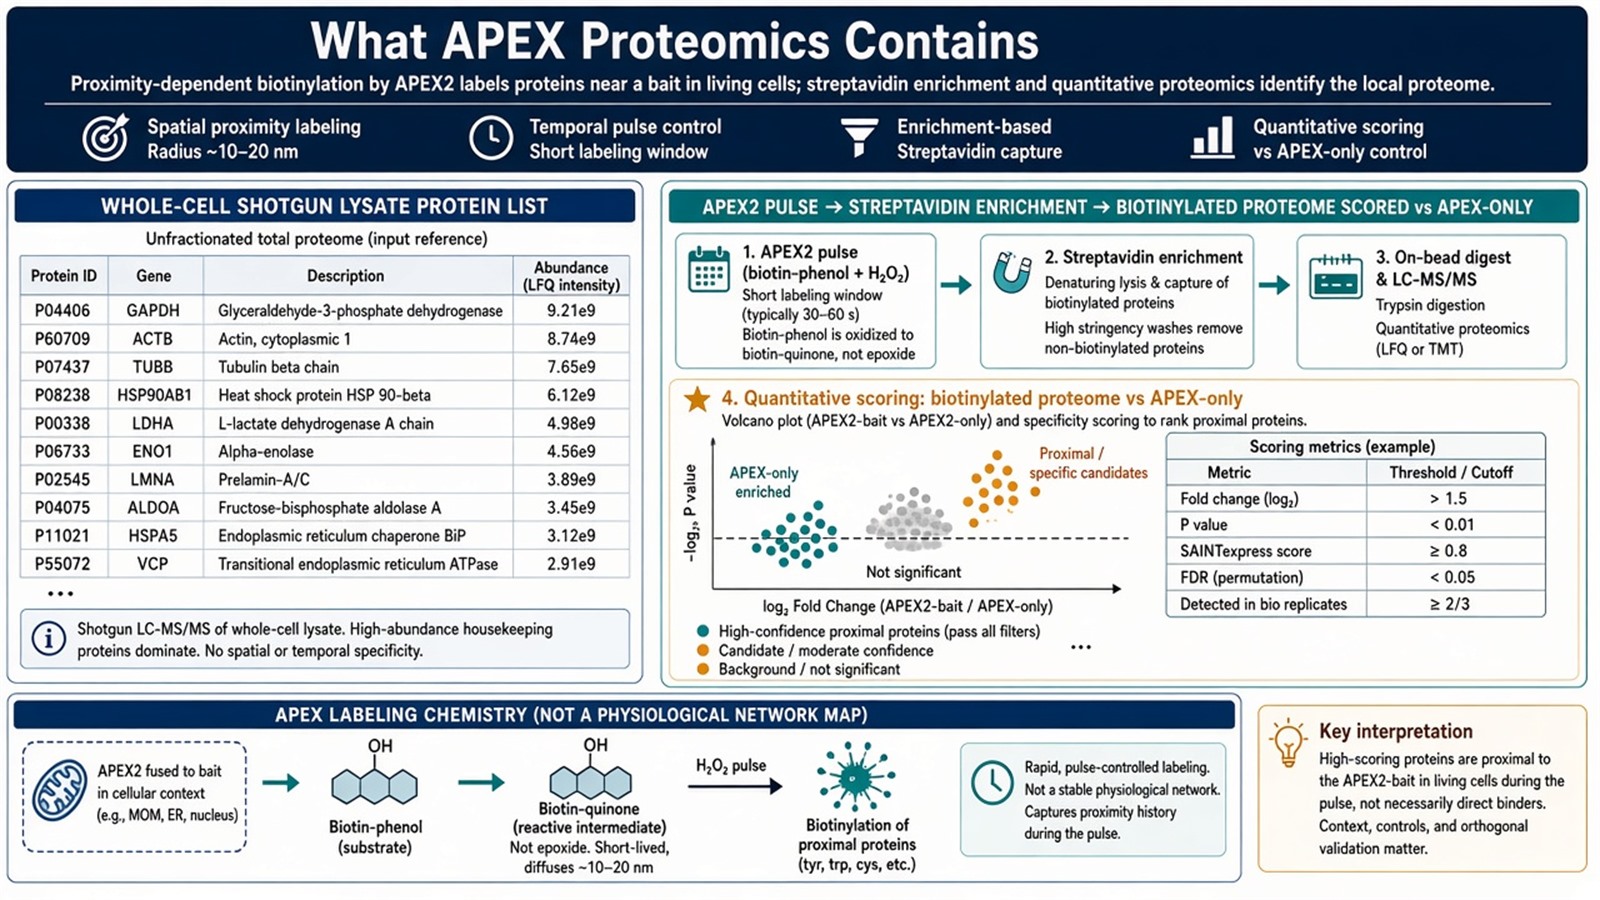
Figure 1. Shotgun proteomics names proteins in a lysate. APEX proteomics names proteins recovered after an APEX2 pulse and streptavidin. APEX-only estimates enzyme and probe background.
Write the APEX-only digest before ranking a protein as localized or interacting. A high peptide count on a matrix protein next to overexpressed APEX2 is still a neighborhood row.
Two Proteomic Jobs
These jobs do not share a control.
A compartment proteome uses APEX2 fused to a targeting peptide. The comparison is a matched APEX2 that is cytosolic or mislocalized. Mitochondrial matrix mapping is the historical example. The output is enrichment of that compartment relative to the control, not a complete organelle inventory.
A bait-neighborhood proteome uses APEX2 fused to a protein of interest. The comparison is localization-matched APEX-only. Transient neighbors can be biotinylated. They are not thereby a signal-transduction mechanism, and the intensities are not high-resolution quantitative proof of a pathway.
Ratiometric labeling, SILAC, TMT, or label-free quantification can compare those arms. The ratio is occupancy of the streptavidin bead after a written pulse. It is not a spatial coordinate and not a binding constant. For the ligase-based neighborhood table without H₂O₂, How Can TurboID-Based Proximity Labeling Enable the Study of Low-Abundance Protein Interactions? provides a complementary TurboID page. For when a partner list versus a residue pair is the next sentence, Co-IP-MS vs XL-MS: Which Protein Interaction Workflow Should You Choose? provides a complementary method choice.
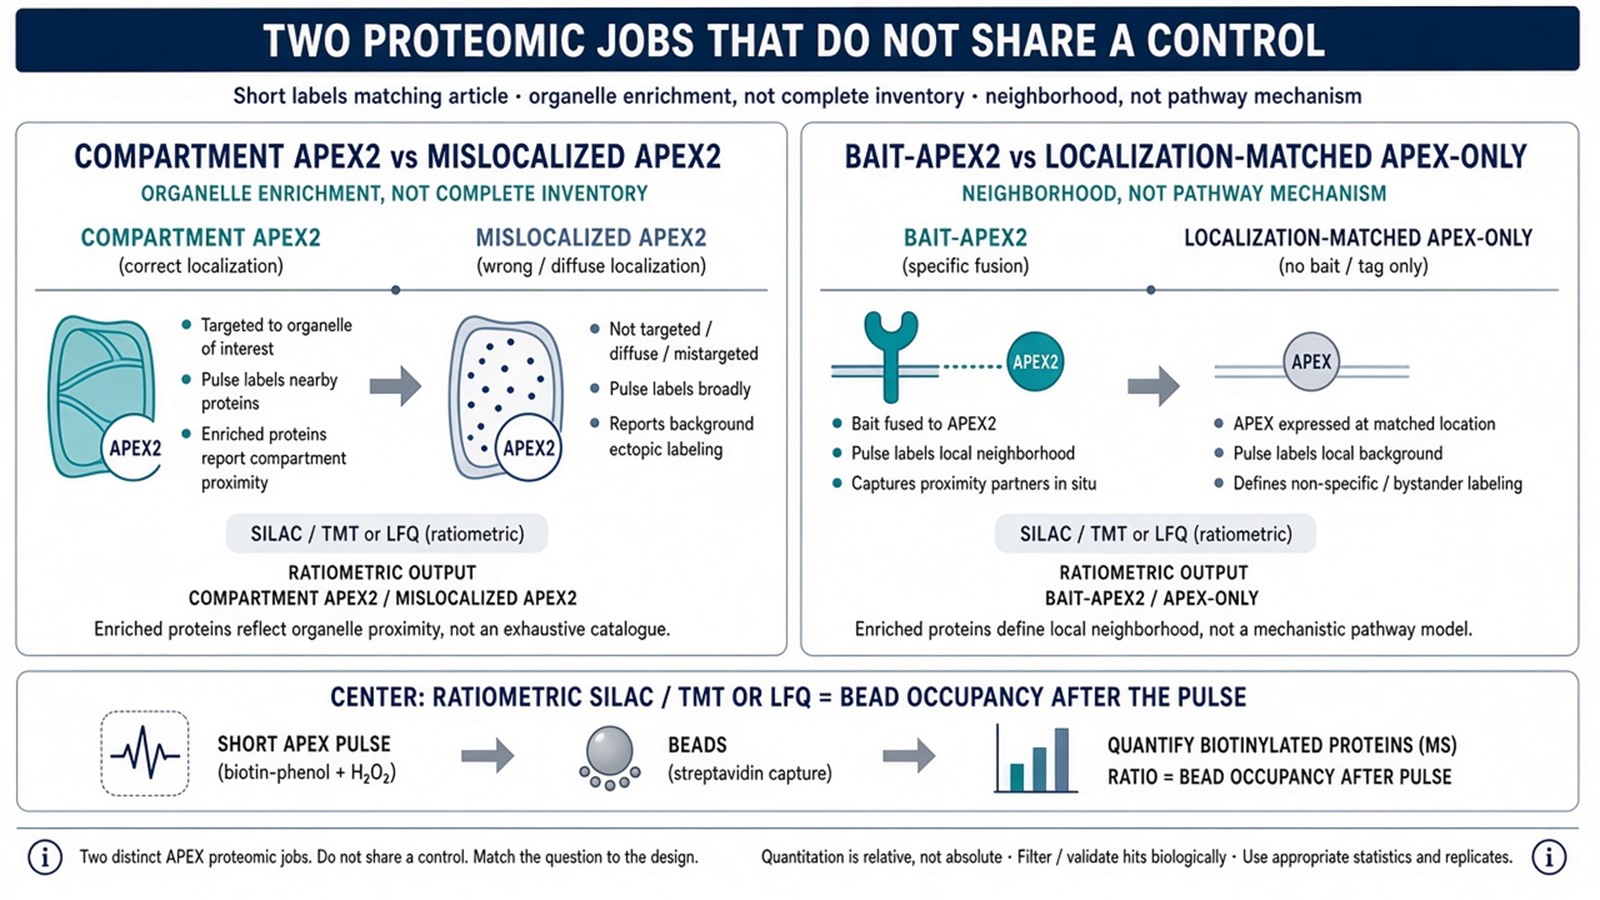
Figure 2. Compartment APEX2 is scored against mislocalized APEX2. Bait–APEX2 is scored against ligase-free or cargo-free APEX2 at the same site. One control does not serve both jobs.
What APEX Proteomics Is Not
Phenolic probes are the standard APEX2 reagents. Epoxide derivatives are not the default APEX proteomic chemistry and should not be listed as if they were. Non-disruptive capture is false after H₂O₂, lysis, and streptavidin.
Drug-target identification requires a compound-centered design, not an APEX2 fused to a suspected target. Reconstructing complex interaction networks from one streptavidin file is a later overlay. Instrument resolution does not make identification and quantification accurate if the arms were unmatched. For how identification names are produced after any pull-down-class enrichment, Mechanism of Pull-Down Coupled with MS for Protein Identification provides a complementary naming discussion.
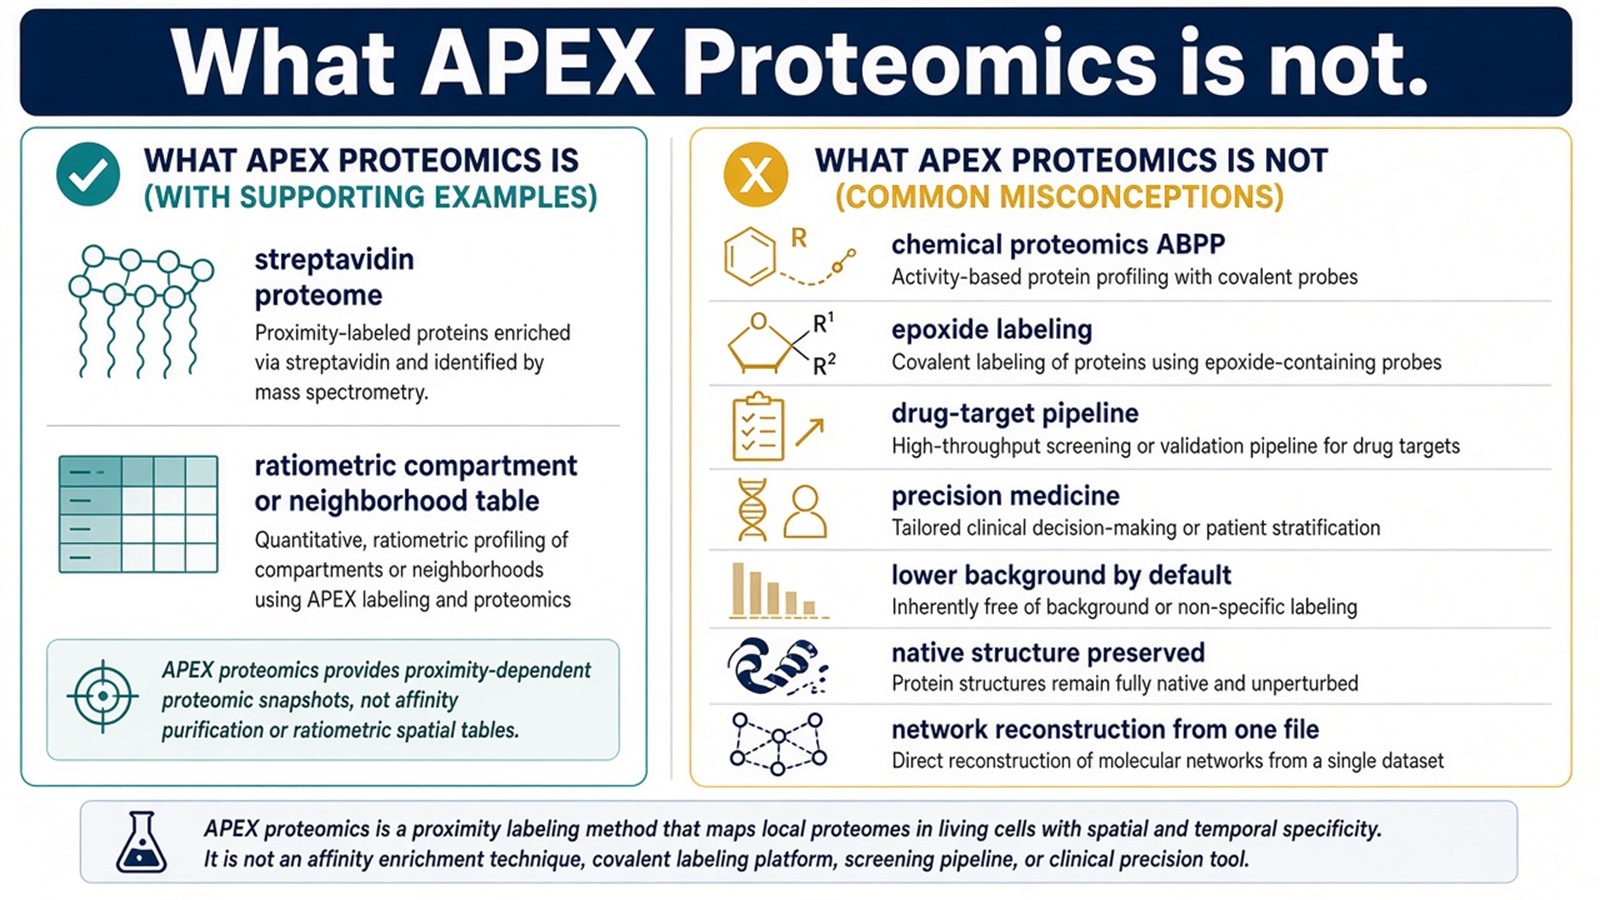
Figure 3. Supported: a ratiometric streptavidin proteome after a defined pulse. Not granted by peroxidase efficiency: lower background, physiological networks, or therapeutic-target validation.
Limits That Sit in the Proteomic Design
Intracellular environment, H₂O₂ access, and tyrosine density change who is labeled. Those are chemistry limits, not only optimization tasks. High-resolution MS is required to name the eluate. It does not rescue a missing APEX-only run or convert the table into dynamics.
Experimental Limits
APEX proteomics reduces a living cell to what a peroxidase fusion, a biotin-phenol/H₂O₂ pulse, streptavidin, and the search engine can report. Diagnostic, precision-medicine, or pipeline language sits outside ordinary research reporting unless a separately validated assay framework is in place.
Project Planning and Service Scope
A usable APEX proteomic plan names the job (compartment or bait), the matched APEX-only arm, the quantitative method, and the claim that will be allowed, before the first pulse. MtoZ Biolabs scopes proximity-labeling MS with Orbitrap-class LC-MS/MS. Interpretability still follows those arms, not a chemical-proteomics slogan.
Once the next sentence is a wash-stable endogenous pellet rather than a streptavidin proteome, the project should not be forced through APEX2. Researchers who need antibody capture can review the MtoZ Biolabs Co-Immunoprecipitation Protein Interaction Analysis Service for project-specific sample evaluation, feasibility assessment, and workflow planning.
Frequently Asked Questions
1. What is APEX proteomics?
LC-MS/MS of proteins biotinylated by APEX2 and recovered on streptavidin. It is a spatial or neighborhood proteome, not a whole-cell interaction map.
2. Does APEX proteomics have lower background than other labeling methods?
Not by itself. Background is estimated by APEX-only and no-pulse arms. Higher peroxidase activity can raise that background.
3. Can APEX proteomics identify drug targets?
Not by fusing APEX2 to a suspected mechanism protein. Target identification is a different design.
4. Are epoxide probes part of standard APEX proteomics?
No. Biotin-phenol is the usual reporter. Epoxides belong to other covalent-chemistry experiments.
5. How does this page differ from the APEX proximity-labeling page?
That page is the peroxidase pulse. This page is how the streptavidin file is designed and read as a proteome.
Reference
- Rhee H.W., Zou P., Udeshi N.D., et al. (2013). Proteomic mapping of mitochondria in living cells via spatially restricted enzymatic tagging. Science, 339, 1328-1331. https://doi.org/10.1126/science.1230593
- Hung V., Zou P., Rhee H.W., et al. (2014). Proteomic mapping of the human mitochondrial intermembrane space in live cells via ratiometric APEX tagging. Mol. Cell, 55, 332-341. https://doi.org/10.1016/j.molcel.2014.06.003
- Hung V., Udeshi N.D., Lam S.S., et al. (2016). Spatially resolved proteomic mapping in living cells with the engineered peroxidase APEX2. Nat. Protoc., 11, 456-475. https://doi.org/10.1038/nprot.2016.018
- Lam S.S., Martell J.D., Kamer K.J., et al. (2015). Directed evolution of APEX2 for electron microscopy and proximity labeling. Nat. Methods, 12, 51-54. https://doi.org/10.1038/nmeth.3179
- Gingras A.C., Abe K.T., Raught B. (2019). Getting to know the neighborhood: using proximity-dependent biotinylation to characterize protein complexes and map organelles. Curr. Opin. Chem. Biol., 48, 44-54. https://doi.org/10.1016/j.cbpa.2018.10.017
