



A tandem mass tag (TMT) workflow turns separately digested samples into one labeled peptide pool, then reads relative protein change from reporter ions in a shared liquid chromatography-tandem mass spectrometry (LC-MS/MS) run. The sequence is not a generic proteomics checklist. Channel loads must be matched before labeling, unused tag must be removed, and the multiplex set must stay closed after mixing. Fractionation and MS2 versus MS3 acquisition change how much co-isolation reaches the reporters, but they cannot repair an unmatched digest. Reporter-ion ratios support relative comparison inside that set. They do not replace a colorimetric total-protein assay, and they do not report protein amounts.
Researchers who already have defined samples, comparison groups, or a specific project objective can also review the MtoZ Biolabs TMT Quantitative Proteomics Analysis Service for project-specific feasibility and analysis planning.
When a TMT Workflow Is the Execution Path
A TMT workflow is the right execution path when the contrast can be frozen inside one multiplex kit, peptides can be labeled at amines, and the scientific claim is a relative ratio across those channels (Thompson et al. 2003). It is the wrong execution path when samples must remain independent injections, when enrollment will continue after mixing, or when the endpoint is a calibrated amount.
Acquisition is a separate decision from labeling. In MtoZ isobaric-labeling offerings, TMT workflows are typically acquired in data-dependent acquisition (DDA) so reporter ions can be read. Data-independent acquisition (DIA) is not a TMT step.
Researchers who need to judge whether those workflow constraints are worth the same-run comparison can refer to Advantages and Disadvantages of TMT-Based Quantitative Proteomics Analysis for the design-level trade-offs.
From Extracted Protein to a Labeled Peptide Pool
Extraction, digestion, and labeling determine whether later reporter ions still represent the intended biological groups. Errors at this stage cannot be removed by pathway software.
Load matching before digestion
Proteins are extracted from cells, tissues, or biofluids under conditions compatible with digestion and amine labeling. A colorimetric reading such as bicinchoninic acid (BCA) or Bradford is used to match protein input across channels, not to quantify individual proteins. Unequal loading becomes an artificial ratio after tags are mixed.
Digestion, usually with trypsin after reduction and alkylation, creates the amines that TMT reagents modify. Incomplete digestion leaves missed-cleavage peptides that do not represent the parent protein evenly across channels. Amine-containing buffers that survive cleanup compete with peptides for reagent and leave unlabeled species in the pool.
Labeling, quenching, and removal of excess tag
Each digest receives one channel from a commercially available TMT kit. The reagent reacts with peptide N-termini and lysine residues. After the reaction is quenched, excess tag is removed, commonly by reversed-phase cleanup, so leftover reagent does not dominate later spectra or occupy chromatographic capacity.
Incomplete labeling, over-labeling of unexpected sites, or residual free tag all change reporter intensities without a corresponding biological change. Channel assignment has to be recorded before mixing. A sample cannot be relabeled into the same pool after the set is combined.
Pooling, Fractionation, and Reporter-Ion Acquisition
Labeled peptides are combined into one mixture. From this point, every channel shares chromatography, ionization, and precursor selection. That shared path is the reason TMT reduces injection-to-injection drift. It is also the reason a failed channel cannot be reacquired without repeating the plex.
The pooled mixture is often fractionated, for example by high-pH reversed-phase chromatography or, in older designs, strong cation exchange, to lower peptide density per LC-MS/MS run. Fractionation can increase identification depth and can reduce co-isolation that compresses reporter ratios (Ting et al. 2011). It also multiplies instrument time and requires the same fraction boundaries to be kept for the whole set.
Each fraction is analyzed by LC-MS/MS. Precursors are isolated and fragmented. Reporter ions of distinct masses appear in the fragment spectrum and are used as the quantitative signal. MS2 reporters are taken from the isolation window and remain vulnerable to co-isolated peptides. Some tribrid methods use synchronous precursor selection MS3 (SPS-MS3) to read reporters from a second fragmentation of selected fragments, which can reduce compression (McAlister et al. 2014). MS3 is an acquisition choice inside the TMT workflow. It is not a substitute for matched loading or complete labeling.
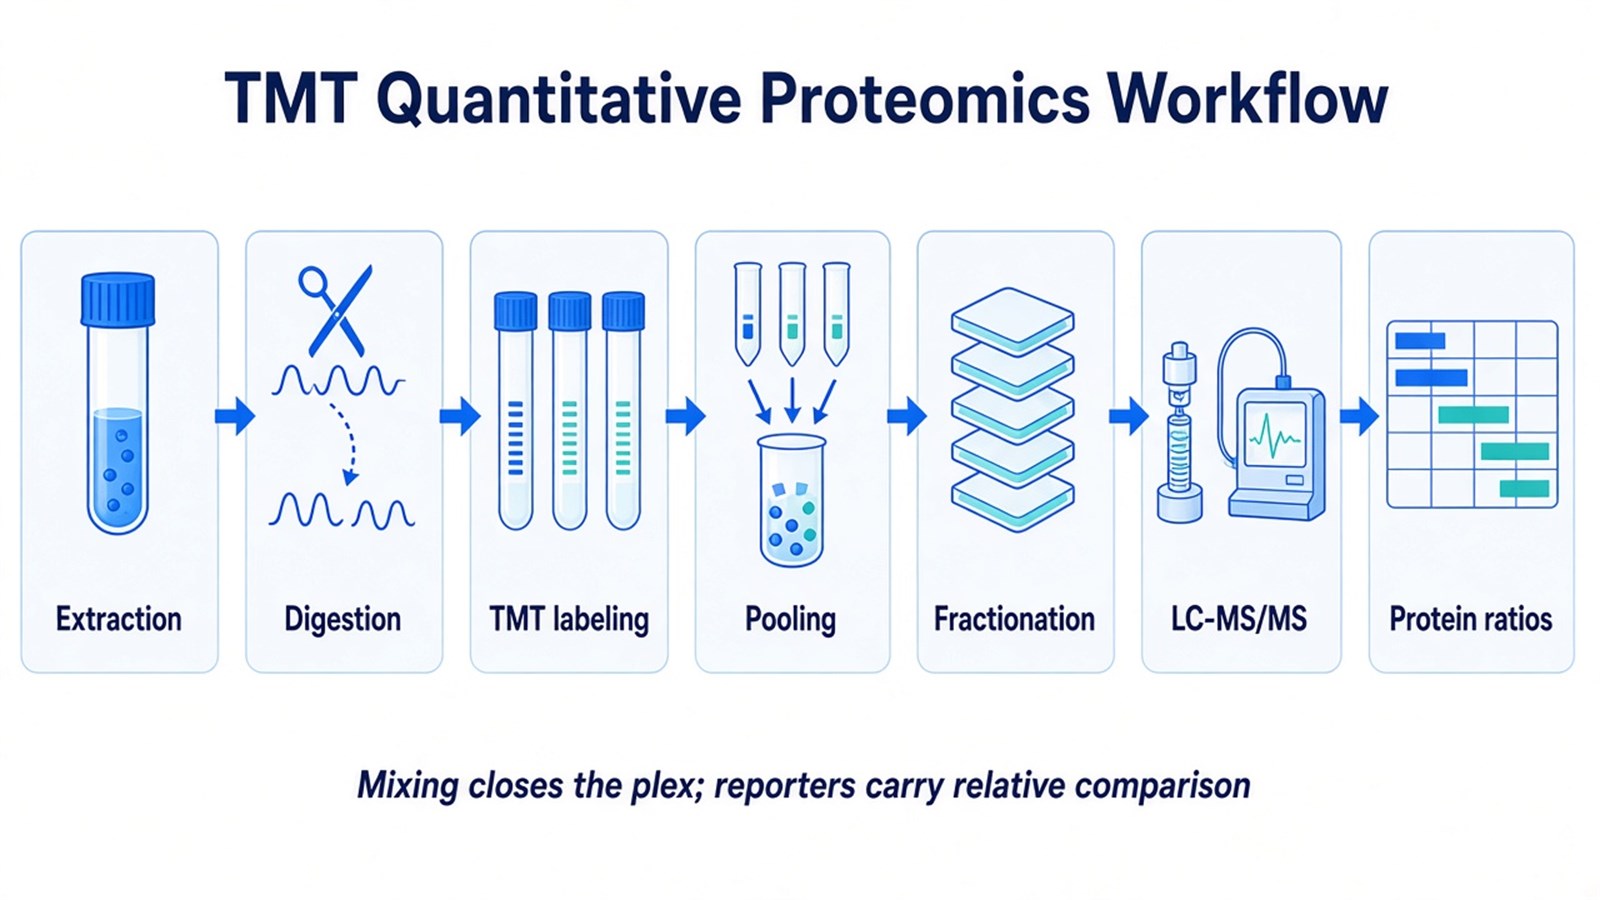
Figure 1. A TMT workflow creates one labeled peptide pool; reporter ions in the shared LC-MS/MS run become protein-level ratios only after identification and channel summarization.
Once samples are mixed, later enrollment is a new plex, not an extra injection in the same ratio matrix.
What Data Processing Must Preserve
Software such as Proteome Discoverer or MaxQuant assigns peptide identifications, extracts reporter intensities, corrects isotope impurities where the method requires it, and summarizes peptides to proteins. The quantitative object remains a relative channel ratio inside the plex.
Normalization can adjust global channel intensity. It cannot repair a confounded labeling batch, a swapped channel map, or a digest that was never fully labeled. Differential testing and pathway annotation are downstream interpretation. They are not TMT workflow steps, and they do not validate a compressed ratio.
Western blot or enzyme-linked immunosorbent assay (ELISA) can test a short candidate list in a new aliquot. Those assays are confirmation experiments with their own antibodies and linearity limits. They are not step 8 of TMT acquisition.
Workflow Control Points That Change the Result
Each stage controls a different failure mode. Skipping the control does not always stop protein identifications from appearing, which is why a complete-looking matrix can still be a weak comparison.
Workflow stage |
What it must keep comparable |
If the control is weak |
|---|---|---|
Extraction and load matching |
Protein input and cleanup across channels |
Ratios track loading, not biology |
Digestion |
Cleavage completeness and amine availability |
Shared or missed-cleavage peptides distort protein values |
TMT labeling and cleanup |
Channel identity, labeling completeness, excess-tag removal |
Unlabeled peptides and leftover reagent mix into reporters |
Pooling |
Closed multiplex membership |
Later samples cannot join the same ratio matrix |
Fractionation |
Peptide density entering each LC run |
Co-isolation and missing depth both increase |
MS2 or MS3 acquisition |
How reporters are isolated |
Fold changes may be compressed toward one |
Protein summarization |
Peptide-to-protein rules and impurity correction |
Protein numbers move without a new spectrum |
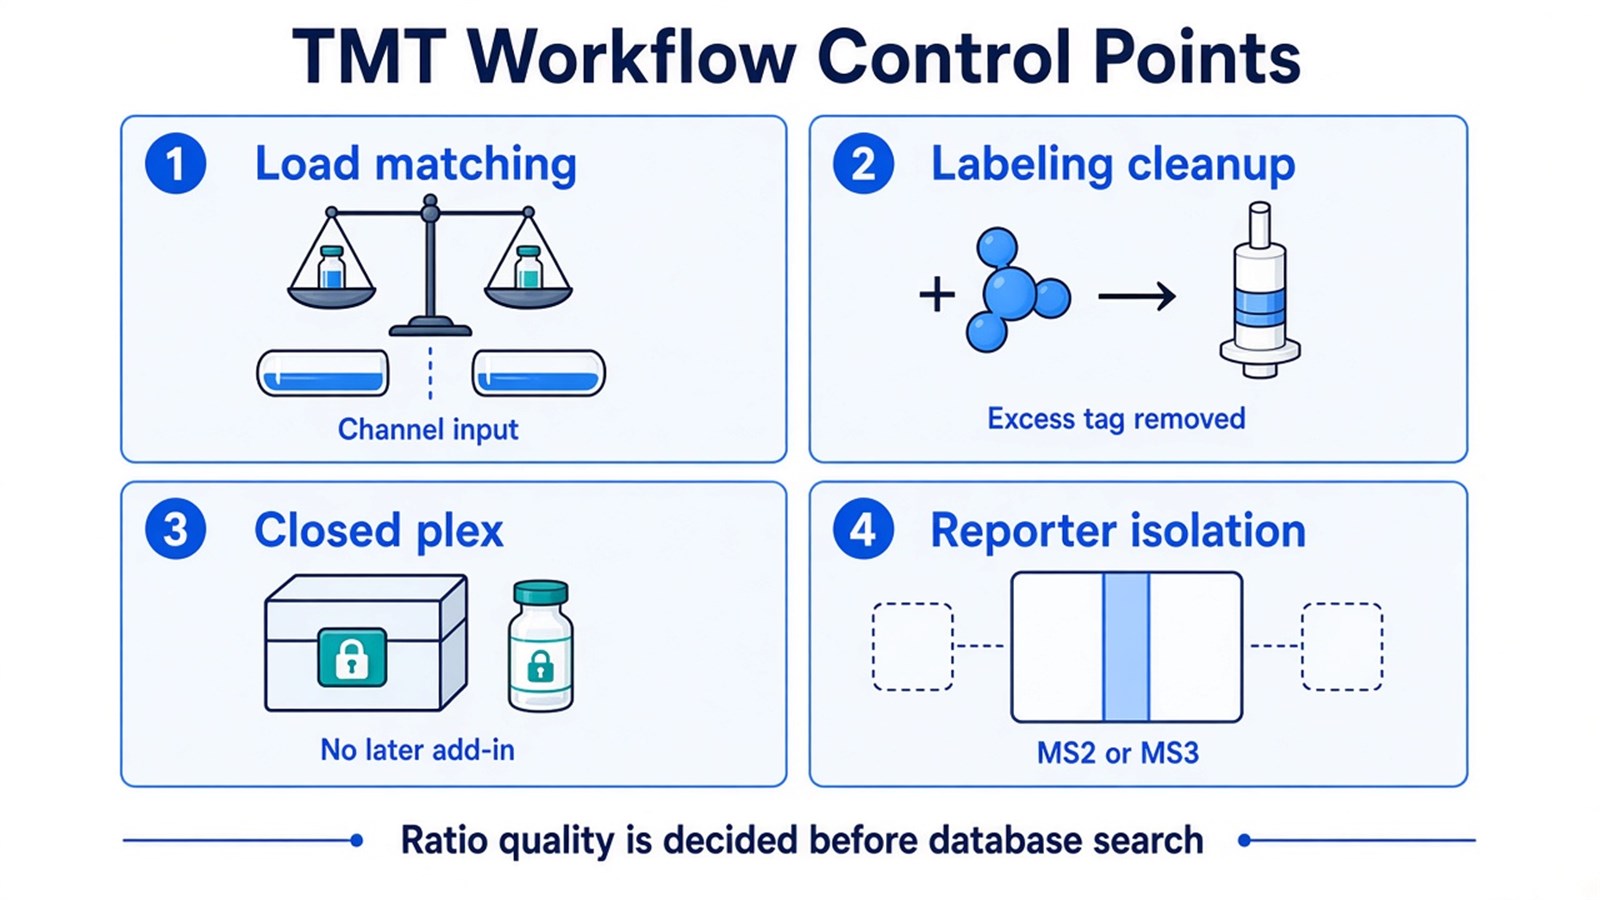
Figure 2. TMT workflow quality is decided before database search: matched loads, complete labeling, a closed plex, and how reporter ions are isolated.
Frequently Asked Questions
1. Why is a BCA or Bradford reading taken before TMT labeling?
The colorimetric assay matches protein input across channels so that later reporter ratios are not dominated by unequal loading. It does not identify proteins and is not a TMT quantification result.
2. Is fractionation required in every TMT workflow?
No. Fractionation is used to lower peptide complexity and can reduce co-isolation in dense samples. A simple plex with limited material may proceed without deep fractionation. The choice trades depth and compression risk against instrument time.
3. Can additional samples be added after the labeled peptides are mixed?
Not into the same ratio matrix. Mixing closes the plex. A later sample requires a new labeling set and is a new analytical block.
4. Does the TMT workflow include Western blot or pathway enrichment?
Those are optional follow-up analyses. The TMT workflow ends at reporter-ion protein ratios, with peptide support and the stated normalization attached. Antibody assays and pathway maps answer different questions.
Once extraction chemistry, channel map, fractionation plan, and expected protein-level readout are defined, the project can be evaluated as a TMT workflow. Researchers can review the MtoZ Biolabs TMT Quantitative Proteomics Analysis Service for project-specific sample evaluation, feasibility assessment, and workflow planning.
