



TMT-based proteomics is a relative quantitative strategy that chemically labels peptides from different samples with isobaric tandem mass tags, mixes those digests, and compares reporter-ion intensities in one liquid chromatography-tandem mass spectrometry (LC-MS/MS) run (Thompson et al. 2003). The tags are indistinguishable at the MS1 level, so the same peptide from every channel co-elutes as one precursor. Fragmentation releases reporter ions of distinct masses, and those intensities become channel ratios, then protein-level values. The valid claim is relative change of the same protein inside one multiplex set. TMT-based proteomics is not a colorimetric total-protein assay, not an intact-mass identity check, and not an absolute concentration method.
Researchers who already have defined samples, comparison groups, or a specific project objective can also review the MtoZ Biolabs TMT Quantitative Proteomics Analysis Service for project-specific feasibility and analysis planning.
What TMT-Based Proteomics Can Answer
The method fits treatment, time-point, or multi-group contrasts in which every sample can enter the same labeling kit and the paper needs a relative ratio rather than a calibrated amount. Because channels share one injection, chromatographic drift between separately acquired unlabeled files is no longer the main comparison noise. Cultured cells, tissues, and biofluids can enter this route when extraction leaves peptides that can be labeled at amines. Metabolic labeling such as SILAC is a different constraint: the organism must incorporate heavy amino acids, which TMT does not require.
It cannot report copies per cell, a tissue concentration, or a diagnostic cutoff. It cannot recover a sample that arrives after the pool is mixed. Disease-name lists do not change those limits. A cancer, neuroscience, or drug-response study still has to pass the same tests: closed groups, labelable peptides, and a relative claim.
Unlabeled discovery and targeted monitoring answer adjacent questions. Unlabeled label-free quantification keeps each digest as its own LC-MS/MS file. Targeted assays such as multiple reaction monitoring (MRM) measure predefined peptides. TMT sits between those needs: multiplexed relative discovery inside one pooled run.
How Isobaric Tags Create a Same-Run Comparison
A TMT reagent has a reactive group that labels peptide N-termini and lysines, a mass-normalizing balancer, and a reporter that is released on fragmentation. The balancer offsets the reporter mass so that tagged peptides from different channels remain isobaric until fragmentation. After digestion, each sample receives one channel from a commercially available kit. Labeled peptides are combined, often fractionated to lower complexity, and acquired by LC-MS/MS. Protein values are inferred from identified peptides, so shared peptides, missed cleavage, and incomplete labeling can move the reported number.
Acquisition is independent of labeling. In MtoZ isobaric-labeling offerings, TMT workflows are typically acquired in data-dependent acquisition (DDA) so reporter ions can be read. Data-independent acquisition (DIA) is not a TMT synonym.
Researchers who need the step-level controls can refer to A Detailed Workflow of TMT-Based Quantitative Proteomics for how each stage changes ratio quality.
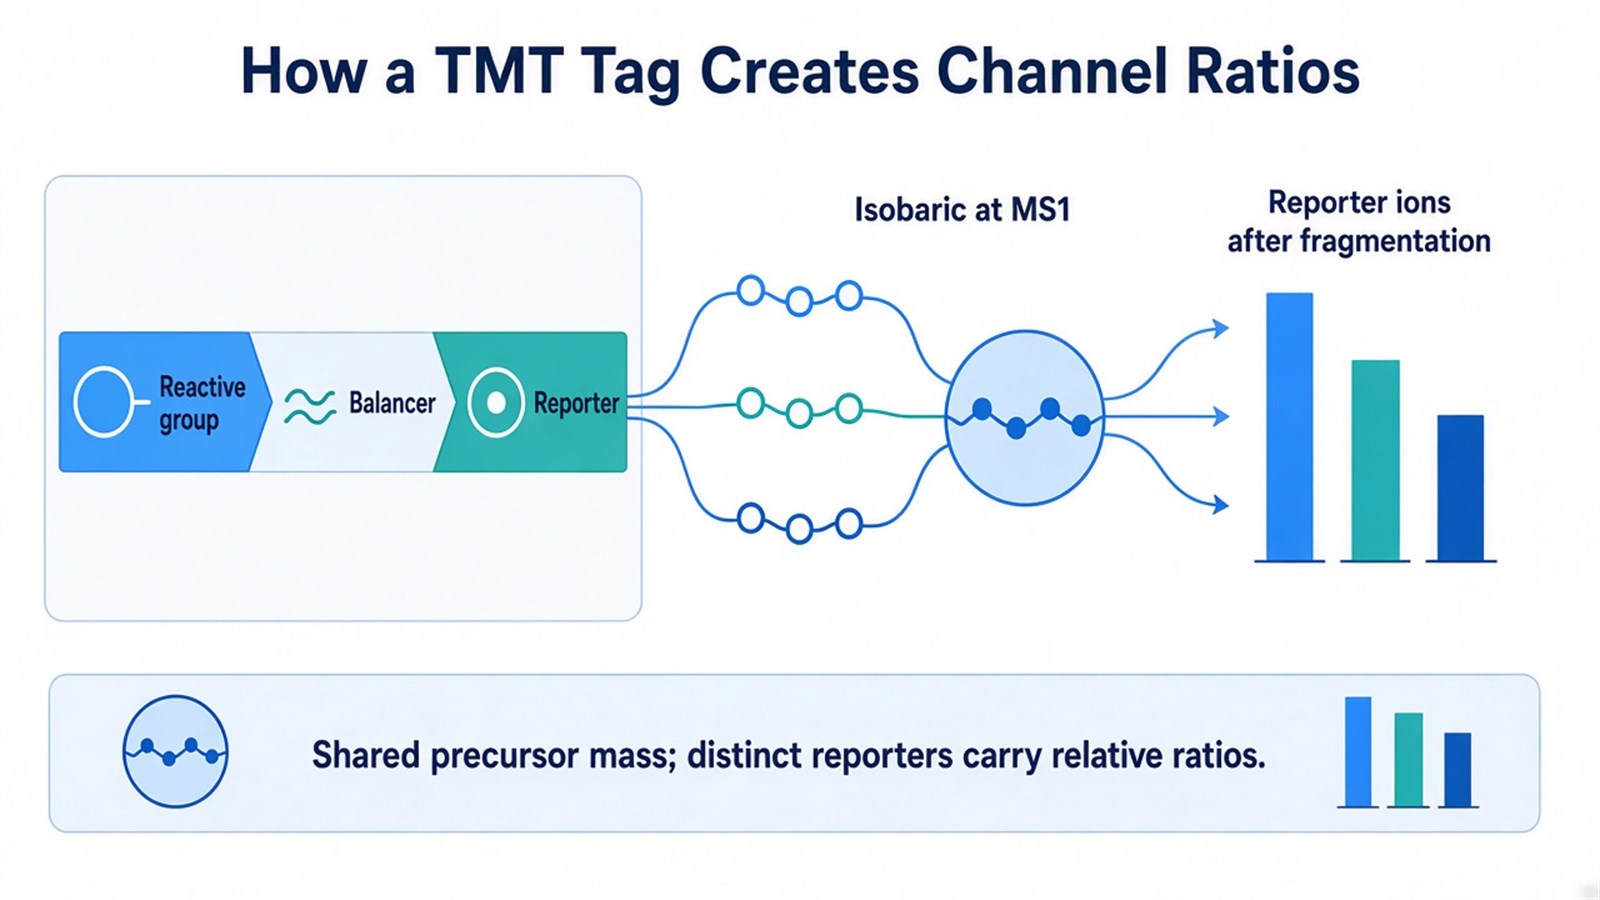
Figure 1. TMT channels share one MS1 precursor mass; reporter ions released on fragmentation carry the relative comparison.
TMT-based proteomics compares channels inside one multiplex set, not independently injected unlabeled files.
When TMT Is the Matching Route
TMT is often the matching route when groups and replicates fit one kit, peptides can be labeled at amines, and same-run relative comparison is the reason for choosing a label. It is a poor match when amine buffers block labeling, when the cohort will keep growing, or when the endpoint is an amount.
Co-isolation in MS2 can compress fold changes toward one (Ting et al. 2011). SPS-MS3 can reduce that mixing on instruments that support the scan (McAlister et al. 2014). Compression is a reason to inspect ratio magnitude, not a reason to treat TMT as uniformly more accurate than unlabeled discovery. A complete quantitative matrix can still hide compressed biology if co-isolated peptides dominate the reporters.
Labeling also consumes peptide material and adds reagent cost. Low-input studies may not yield a complete, balanced plex. Those trade-offs decide whether the workflow should start at all. They do not make TMT a default upgrade over unlabeled discovery for every cohort.
When sample number, multiplexing, or later enrollment makes the choice less straightforward, researchers can refer to Advantages and Disadvantages of TMT-Based Quantitative Proteomics Analysis for when the same-run advantage holds and when unlabeled or targeted routes fit better.
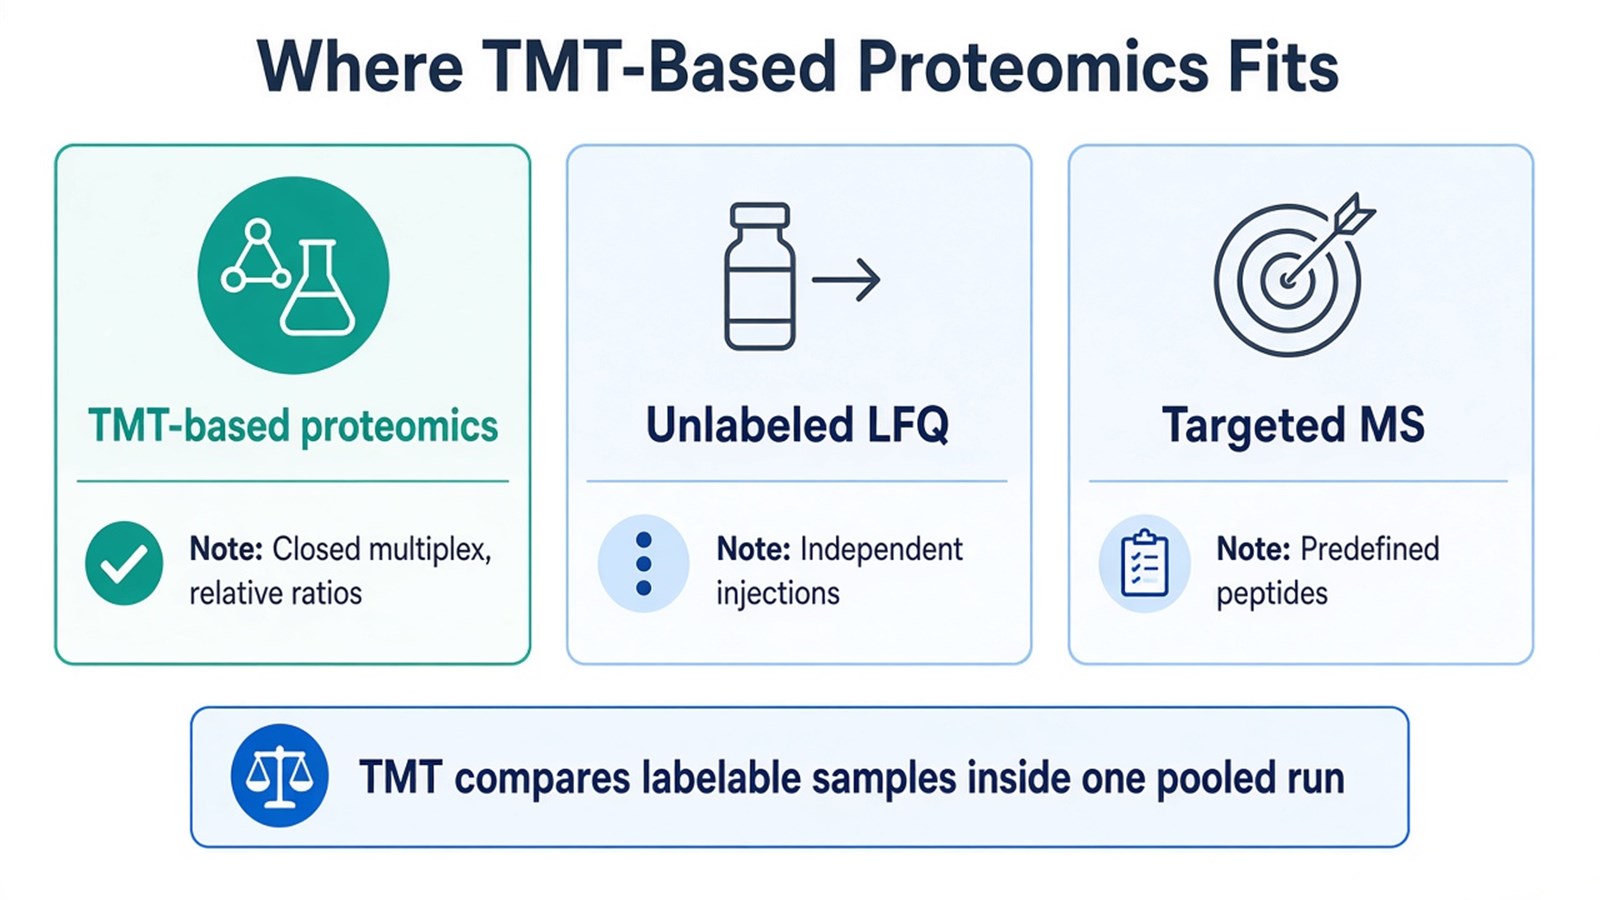
Figure 2. TMT-based proteomics fits a closed, labelable contrast that needs relative ratios from one pooled run.
What a TMT Result Can Support
After identification, reporter extraction, and protein summarization, a higher protein value can support a relative statement about that protein across channels in this plex. Peptide support, missing reporters, impurity correction, and the stated normalization belong next to that statement. A large ratio on one poorly shared peptide is weaker evidence than a smaller ratio supported by several unique peptides.
Ordinary TMT ratios should not rank different proteins by copy number, transfer a fold change from one kit or instrument to another study as a calibrated quantity, or stand in for antibody confirmation. Western blot or ELISA can test a short candidate list in a new aliquot. They are not TMT readout.
Pathway maps and disease labels are interpretation after the ratio table exists. They do not prove that labeling was complete or that compression was negligible.
A TMT protein value is a relative reporter-ion ratio inside one coherent plex.
Frequently Asked Questions
1. Is TMT-based proteomics the same as TMT quantitative proteomics?
The names point to the same labeling strategy. Search titles such as TMT quantitative proteomics and TMT-based quantitative proteomics still describe isobaric tagging, pooling, and reporter-ion relative quantification. They do not name a different acquisition mode or an absolute assay.
2. Does TMT-based proteomics replace Western blot or ELISA?
No. TMT compares many proteins as relative channel ratios. Antibody assays test selected proteins with different specificity and linearity limits. One does not validate the other automatically.
3. Can TMT report absolute protein concentration?
Not from ordinary reporter-ion tables. Amount claims need a matched standard and a calibration model, usually in a targeted experiment.
4. Should a growing cohort be run as TMT-based proteomics?
Only if the current contrast can be frozen inside one kit. Samples that arrive after mixing require a new plex and are a new analytical block. Ordinary normalization does not insert a late digest into the original reporter-ion matrix.
Once the comparison design, labeling feasibility, and expected protein-level readout are defined, the project can be evaluated as a TMT study. Researchers can review the MtoZ Biolabs TMT Quantitative Proteomics Analysis Service for project-specific sample evaluation, feasibility assessment, and workflow planning.
