



Targeted proteomics is operated as a closed-list workflow: select proteotypic peptides for predefined proteins, digest and clean up matched samples, optionally spike stable isotope-labeled standards (SIS), schedule those precursors on a parallel reaction monitoring (PRM) or multiple reaction monitoring (MRM) method, acquire only the listed targets by liquid chromatography-tandem mass spectrometry (LC-MS/MS), and quantify from extracted peak areas. Unlike data-dependent acquisition (DDA) or data-independent acquisition (DIA) discovery, proteins outside the inclusion list remain invisible. Relative same-peptide comparison is the default claim. Absolute amount requires matched SIS peptides and a stated calibration model. The workflow answers confirmation and monitoring questions for a locked panel, not open proteome surveying.
Researchers who already have a closed peptide panel or a validation objective can also review the MtoZ Biolabs Parallel Reaction Monitoring (PRM) Service and SRM-MRM Quantitative Proteomics Services for project-specific feasibility and assay planning.
Targeted Proteomics Workflow Step by Step
A workable targeted project follows a fixed order. Skipping peptide closure or transition validation cannot be repaired by higher sample load alone.
1. Close the protein and peptide list
Define targets from discovery data, literature, or pathway panels. Prefer unique proteotypic peptides with usable chromatography and fragment ions. Shared peptides report gene-family ambiguity rather than a single protein. Flag modification-prone sites when the claim is unmodified abundance.
2. Decide relative versus absolute readout
Same-peptide peak areas support relative confirmation when digestion and loading stay comparable. Absolute concentration needs matched SIS peptides, with spike timing recorded. An SIS added after failed digestion cannot reconstruct the lost protein. SIS is optional for relative targeted work; it is required for absolute claims.
3. Prepare matched digests
Extract, digest, and clean up samples under one protocol. Keep loading matched across the cohort. Matrix-specific lysis and desalting quality bound later peak shapes.
4. Build the PRM or MRM method
Choose the acquisition class for the closed list. PRM isolates scheduled precursors and records full high-resolution MS2 for post-run fragment choice (Peterson et al. 2012). MRM monitors predefined precursor-to-product transitions on a triple quadrupole for high-multiplex production panels (Lange et al. 2008). Optimize retention-time windows, collision energy, and qualifier ions before the cohort is queued.
5. Acquire with QC controls
Run samples, calibrators when used, and QC or pooled injections under the locked schedule. Review retention-time drift, qualifier ratios, and interference before accepting peaks.
6. Integrate peaks and report the stated claim
Extract quantitative and qualifier chromatograms in software such as Skyline. Report relative or SIS-based absolute values only for peptides that pass peak-shape and interference checks. Statistical contrasts and pathway maps are downstream interpretation. They do not convert a closed list into discovery coverage.
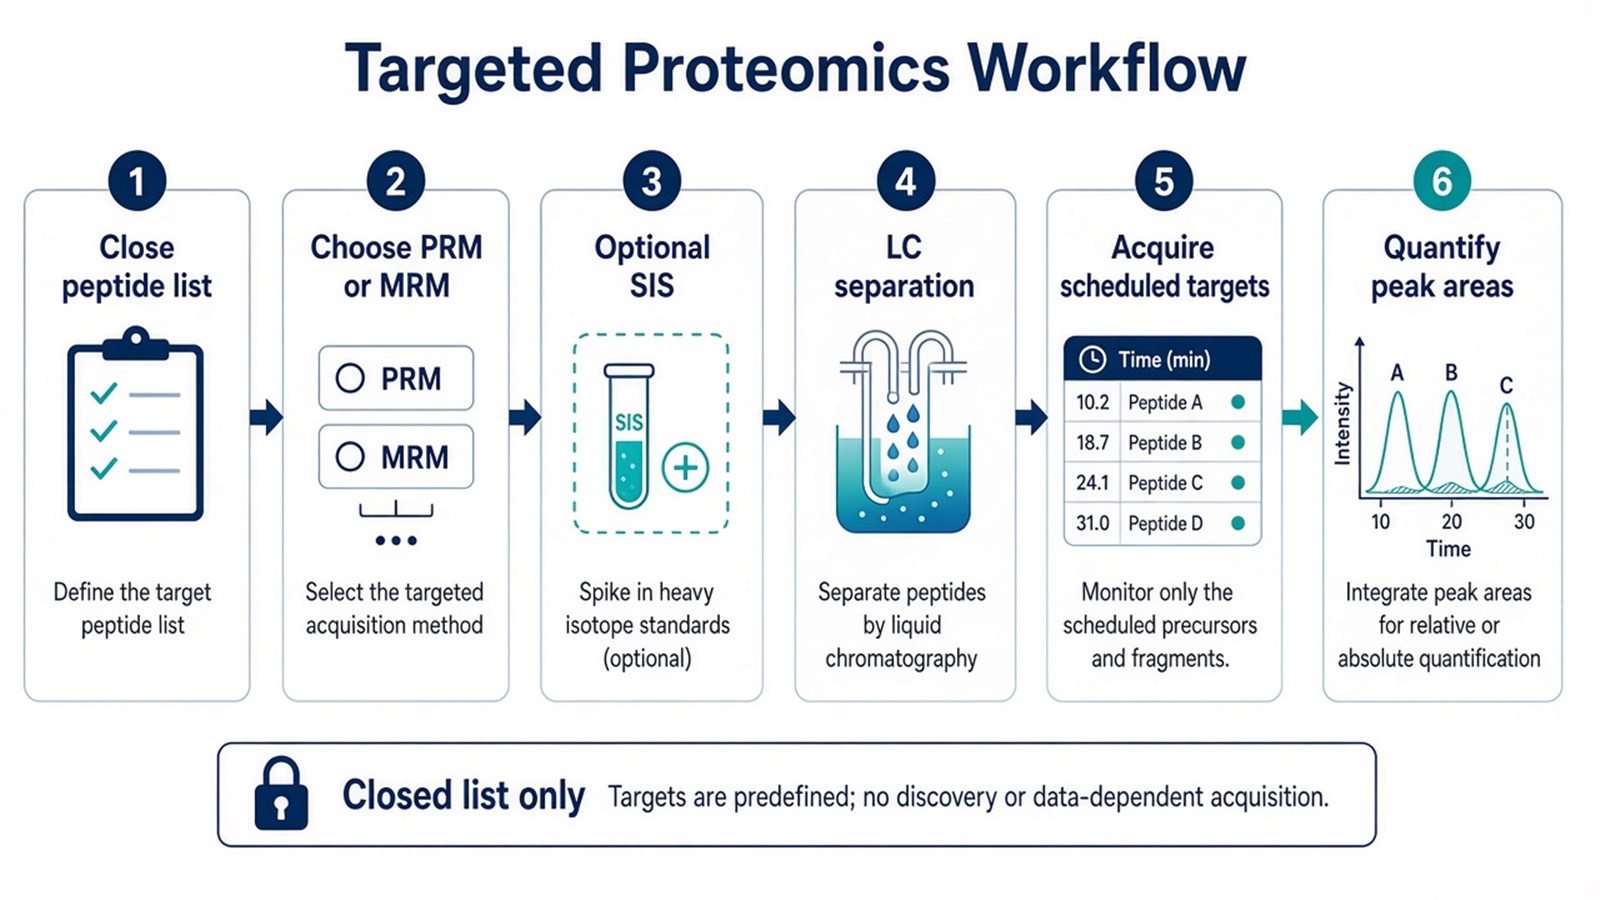
Figure 1. Targeted proteomics is executed as a closed-list workflow; only scheduled peptides are measured.
If the peptide was never placed on the method, targeted acquisition will not measure it, regardless of abundance in the digest.
PRM and MRM Inside the Same Workflow
Both platforms serve the same operational claim: quantify a closed peptide list. They differ in what is recorded and how multiplex is traded against fragment review.
Workflow need |
Prefer PRM |
Prefer MRM / SRM |
|---|---|---|
Method maturity |
Panel still under fragment review |
Transitions already locked |
Matrix |
Complex digests needing accurate-mass inspection |
Mature panel with clean unit-resolution transitions |
Multiplex priority |
Moderate panel with post-run ion choice |
Very high multiplex on a triple quadrupole |
Absolute design |
SIS plus calibration supported on either route |
SIS plus calibration supported on either route |
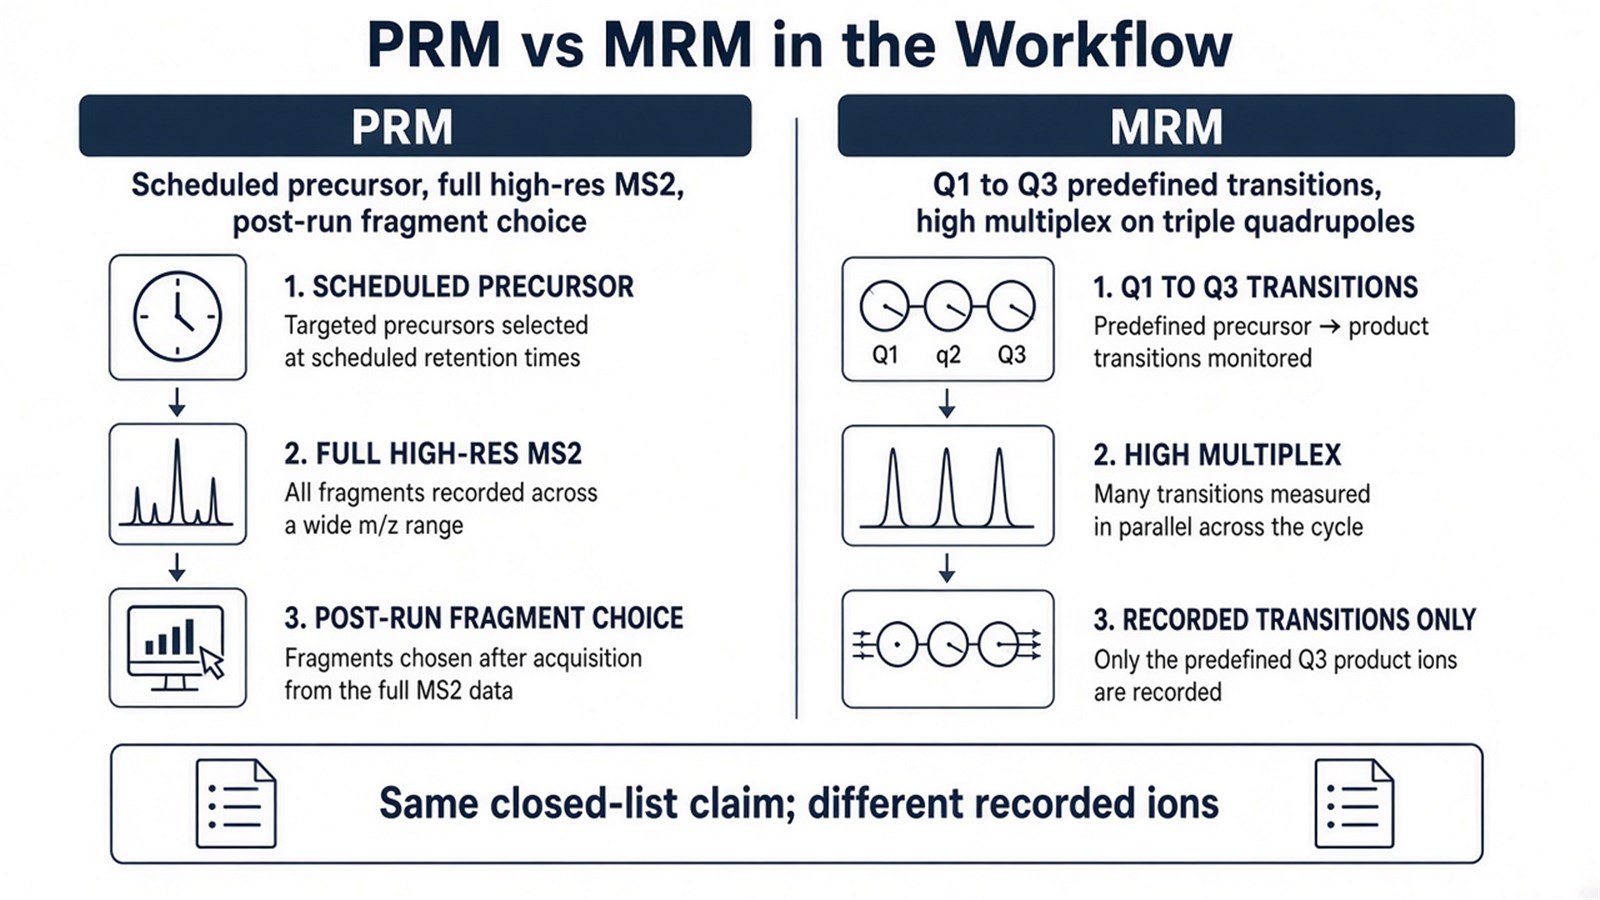
Figure 2. PRM and MRM share the closed-list workflow; they differ mainly in recorded ions and multiplex versus fragment-review trade-offs.
Researchers comparing those platforms can refer to PRM vs. MRM: Comparative Analysis in Quantitative Proteomics. For PRM operational detail, see Parallel Reaction Monitoring (PRM): Principles, Workflow, and Applications. For MRM principle detail, see Principle of Multiple Reaction Monitoring.
Judgment Criteria: When the Targeted Workflow Fits
Study need |
Targeted workflow often fits |
Reassess before scheduling |
|---|---|---|
Target status |
Closed, mature peptide list |
Open discovery inventory still required |
Claim |
Relative confirmation; absolute with SIS |
Clinical diagnostic cutoff without a validated assay framework |
Evidence need |
Fragment- or transition-level monitoring of known targets |
Global proteome ranking |
Upstream path |
DDA/DIA discovery already nominated candidates |
No discovery support and no literature panel |
Scale |
Panel fits PRM cycle time or MRM dwell budget |
Panel too large for the chosen instrument class |
Prefer targeted proteomics when candidates are closed and need confirmation or monitoring. Prefer DDA or DIA while the protein inventory is still open. Prefer PRM when fragment review matters; prefer MRM when a locked high-multiplex transition panel must run repeatedly.
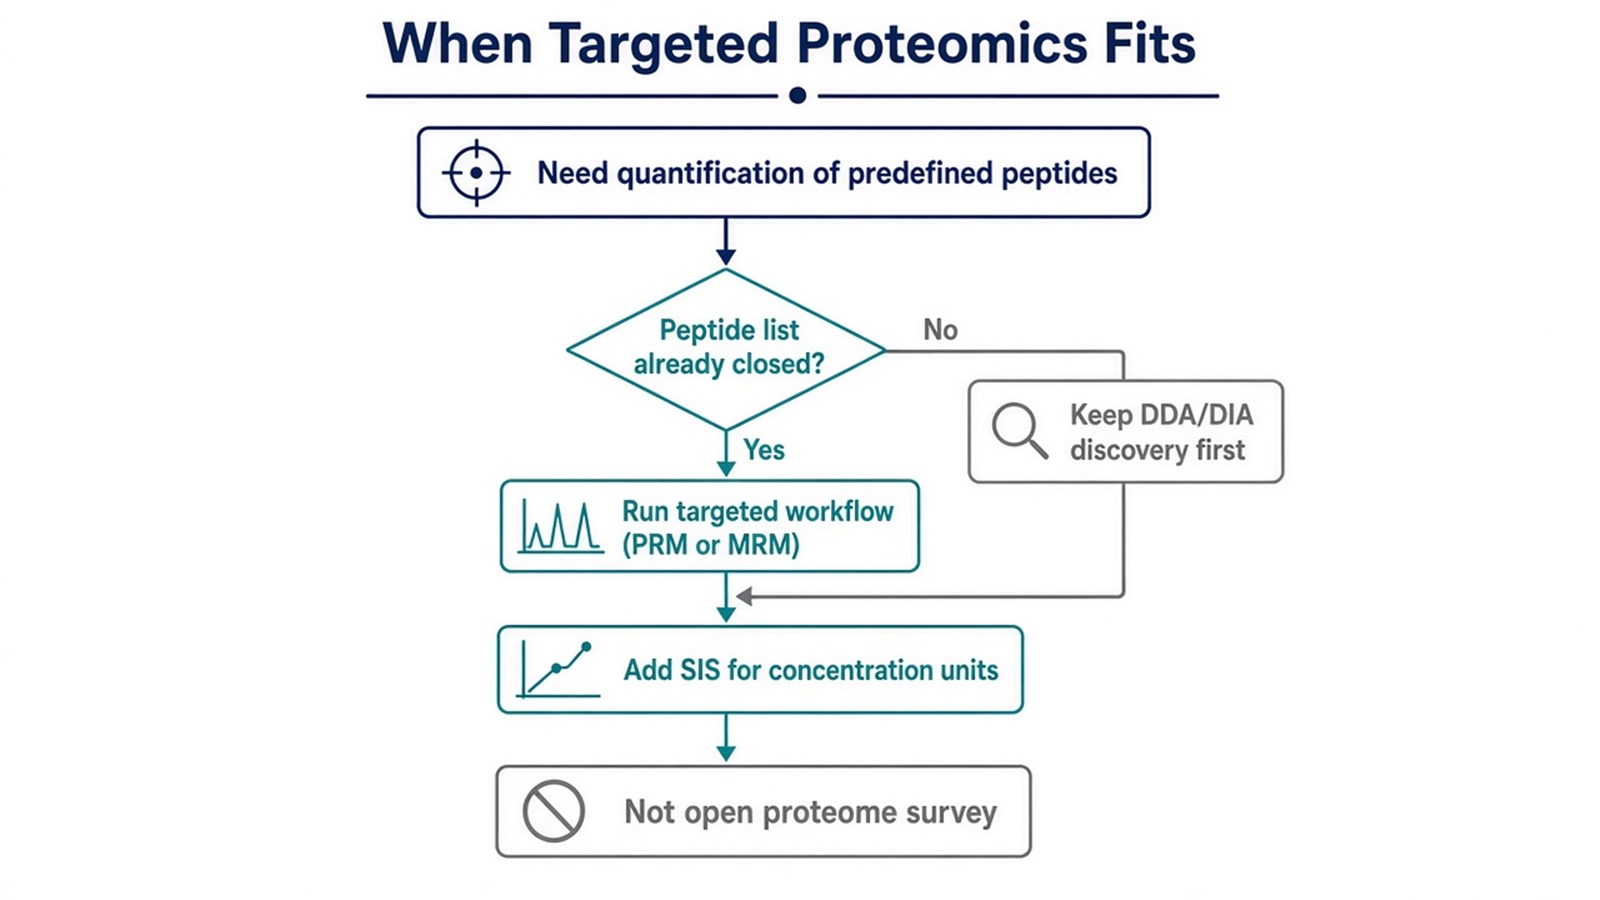
Figure 3. Run the targeted workflow after the peptide list is closed; keep discovery first, and add SIS only when concentration units are required.
Limits and Application Boundaries
Targeted methods cannot recover proteins that were never scheduled. Co-isolation and unit-resolution interference can still admit background ions. Calling every targeted result clinically diagnostic overstates ordinary research reporting. Diagnostic cutoffs and companion-diagnostic claims sit outside this workflow unless a separately validated assay framework is in place.
Typical applications are biomarker candidate validation, pathway-protein panels, pharmacodynamic monitoring, and selected modified-peptide assays when site-discriminating ions can be defined. A common sequence is discovery by DDA or DIA, optional PRM confirmation, then MRM for a locked panel that must run at scale.
Once the peptide list, PRM-versus-MRM choice, and relative-versus-absolute design are defined, the project can be evaluated as a targeted proteomics study. Researchers can review the MtoZ Biolabs Parallel Reaction Monitoring (PRM) Service and SRM-MRM Quantitative Proteomics Services for sample evaluation, method feasibility, and workflow planning. For biomarker-confirmation framing, see How PRM Proteomics Supports Biomarker Validation?.
Frequently Asked Questions
1. How does targeted proteomics work in practice?
It locks a peptide list first, acquires only those scheduled targets by PRM or MRM, and quantifies from peak areas, with SIS required only for absolute units.
2. Is SIS required in every targeted workflow?
No. Relative confirmation is the default without SIS. Absolute amount requires matched SIS and a calibration model.
3. Can targeted proteomics replace DDA or DIA discovery?
No. Targeted methods measure only scheduled peptides. Keep discovery while new protein names are still needed.
4. When is PRM preferred over MRM in this workflow?
When the panel is still under fragment review or accurate-mass discrimination matters in a complex matrix.
5. Does a successful targeted assay equal a clinical diagnostic test?
No. Research confirmation supports a quantitative claim for the studied cohorts. Clinical use needs separate analytical and clinical validation frameworks.
Reference
- A.C. Peterson, J.D. Russell, D.J. Bailey, M.S. Westphall, J.J. Coon (2012). Parallel reaction monitoring for high resolution and high mass accuracy quantitative, targeted proteomics. Mol. Cell. Proteomics, 11, 1475-1488. https://doi.org/10.1074/mcp.O112.020131
- V. Lange, P. Picotti, B. Domon, R. Aebersold (2008). Selected reaction monitoring for quantitative proteomics: a tutorial. Mol. Syst. Biol., 4, 222. https://doi.org/10.1038/msb.2008.61
