



Targeted proteomics analysis starts with the sample record, not with an acquisition preference. Multiple reaction monitoring (MRM) and parallel reaction monitoring (PRM) remeasure a closed peptide panel on liquid chromatography-tandem mass spectrometry (LC-MS/MS) only after matrix, pretreatment, load, and digest rules are matched across every contrast arm (Lange et al. 2008; Peterson et al. 2012). Unmatched extraction or digest shifts the peptide pool and can look like biology after peak integration. Plasma, tissue, lysate, and formulation each need a declared cleanup path before transitions or fragment rules are frozen (Bantscheff et al. 2012).
Relative fold-change and absolute amount remain separate claim units on the same sample set. Absolute reporting needs stable isotope-labeled internal standard (SIS) peptides or Absolute QUAntification (AQUA) design with spike timing aligned to the prep step being corrected (Gerber et al. 2003; Carr et al. 2014). Surrogate peptides report sequences that survive the declared digest. They do not invent every isoform or modified form by default.
In MtoZ Biolabs targeted offerings, targeted proteomics analysis is scoped when matrix type, pretreatment parity, closed panel status, cohort plus quality control (QC) count, and claim unit are written in the brief. Service intake appears in Targeted Proteomics Service.
What Sample and Pretreatment Must Cover
Four sample layers belong in the brief and stay separated.
Matrix layer names plasma, serum, tissue, cell lysate, media, or formulation, plus storage and freeze-thaw history. Each matrix changes suppression and co-elution risk on the same peptide.
Pretreatment layer names extraction, depletion or enrichment if used, reduction, alkylation, digest enzyme, cleanup, and reconstitution. Rules must match on every contrast arm.
Load layer names protein or peptide mass on column from a matrix pilot. Overload compresses dynamic range. Underload drops low-response targets.
Cohort layer names biological replicates, technical repeats if any, blanks, QC pools, and calibration or SIS samples. Study n alone is not the injection count.
Targeted analysis measures peptides that survive a declared prep. Prep drift mid cohort splits the assay record.
How Matrix and Prep Change MRM or PRM Performance
Sample handling and method build are one planning object.
Proteotypic peptide choice depends on digest completeness and uniqueness after the declared enzyme and missed-cleavage policy. A peptide that appears in discovery files may fail after depletion, detergent cleanup, or tissue homogenization.
MRM transition optimization and PRM fragment review must be checked on matrix-matched digests, not only on neat standards (Lange et al. 2008; Peterson et al. 2012). Co-eluting interference tracks cleanup quality and load. Residual lipids or polymers from incomplete prep lengthen method rescue more than adding dwell time.
SIS spike timing must match the claim and the prep step. Peptide standards added after digest do not correct unmatched extraction (Gerber et al. 2003). Enrichment or depletion used on only one contrast arm creates bait-only artifacts that no acquisition mode can repair.
Platform choice after prep is stable belongs on Parallel Reaction Monitoring vs MRM: Which Fits?. Discovery-to-targeted staging when the list is still open appears in Targeted Mass Spectrometry or Discovery LC-MS/MS? Matching MRM, PRM, and Profiling Routes to Your Analytical Goal. Comprehensive targeted MS framing appears in Targeted Mass Spectrometry.
End-to-End Sample-First Workflow for Targeted Analysis
A practical sample-first spine usually runs as follows.
- Freeze contrast, claim unit, and closed peptide priority tiers.
- Document matrix, storage, and pretreatment route for every arm.
- Pilot digest and load on representative material before shipping the full cohort.
- Optimize MRM transitions or PRM fragment rules on matrix-matched digests.
- Size biological n plus QC, blanks, and SIS or calibration injections.
- Lock one prep method and one acquisition method version for the cohort block.
- Acquire interleaved study and QC samples when cohort size allows.
- Integrate peaks; flag prep or target failures against prespecified rules.
- Deliver tables with prep record, method notes, and QC flags; archive methods with the report.
Steps 1 through 3 are sample and prep gates. Steps 4 through 6 lock the assay. Steps 7 through 9 produce the quantitative record. MRM/SRM service context appears in SRM-MRM Quantitative Proteomics Services. PRM service context appears in Parallel Reaction Monitoring (PRM) Service. Absolute reporting intake appears in Absolute Quantitative Analysis (AQUA) Service.
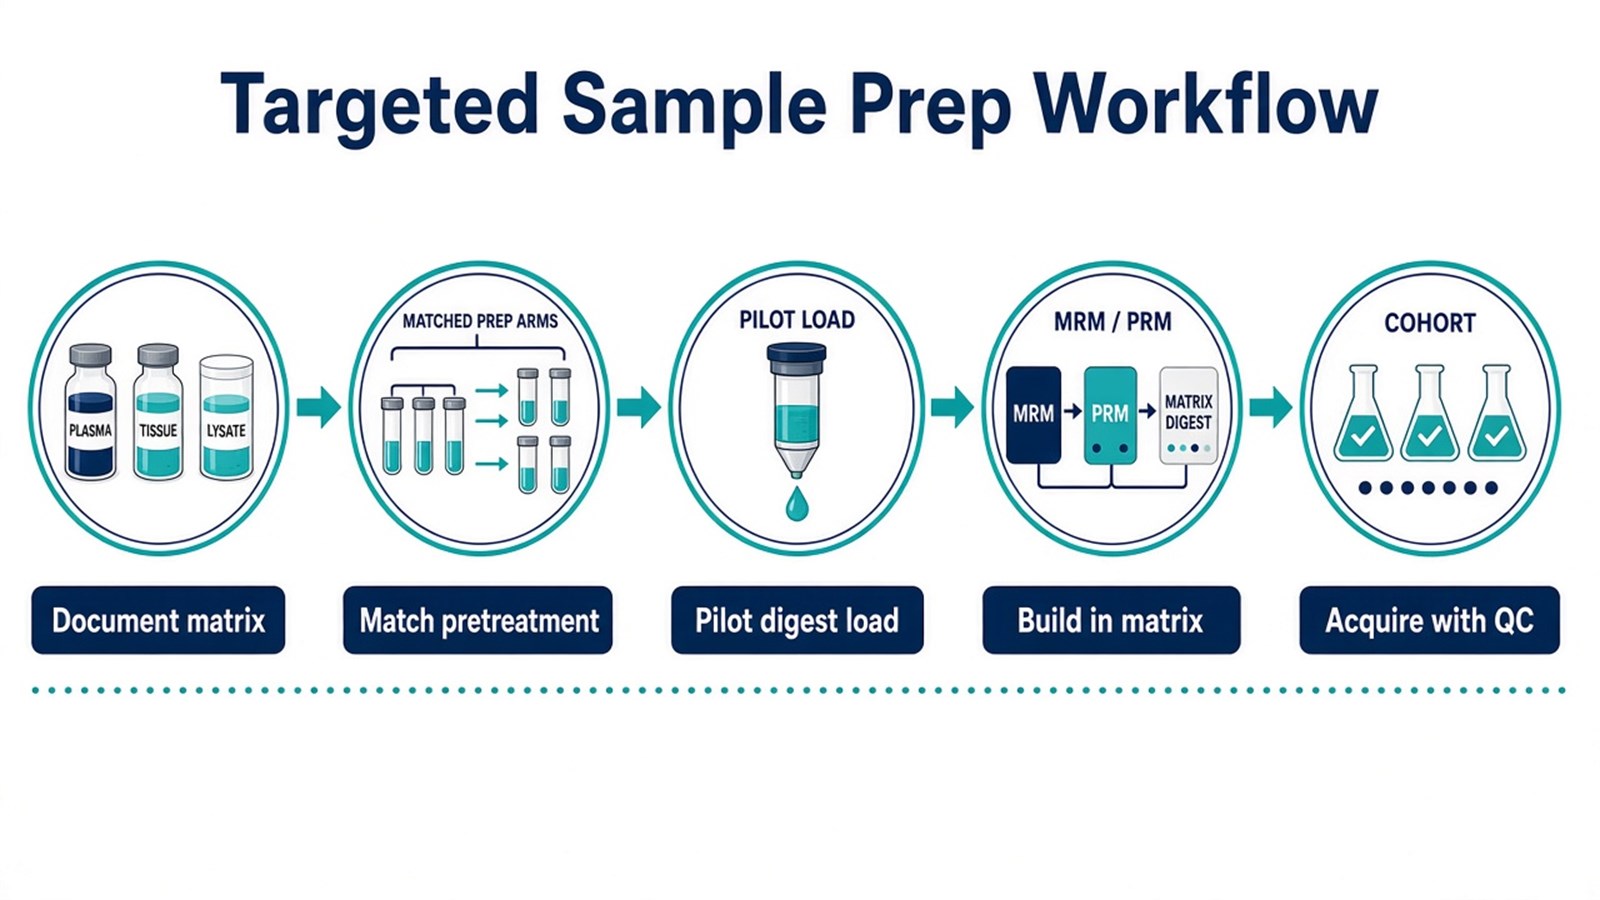
Figure 1. Targeted proteomics analysis is set after matched pretreatment and a matrix-matched pilot; sample count includes QC and calibration injections.
Parameters That Must Be Locked for Sample and Prep
Layer |
Parameter that must be declared |
Failure mode if ignored |
|---|---|---|
Matrix |
Type, storage, freeze-thaw count |
Hidden suppression between arms |
Pretreatment |
Extraction, digest, cleanup parity |
Peptide-pool shift as biology |
Enrichment |
Depletion or pull-down on all arms |
Bait-only artifacts |
Load |
Mass on column from pilot |
Saturation or dropped targets |
Panel |
Peptide list version and priority tiers |
Late growth breaks cycle time |
Platform |
MRM, PRM, or hybrid built in matrix |
Neat-standard methods fail in study matrix |
SIS |
Spike step aligned to claim |
Peptide amount misread as extract amount |
Cohort |
Study n plus QC, blanks, calibrators |
Understated injection load |
Contrast |
Frozen before peak review |
Post hoc arm relabeling |
Deliverable |
Table, prep record, QC depth |
Report rebuilt after acquisition |
Limits follow the same table. Targeted acquisition does not correct unmatched harvest or digest. Large panels do not replace a small, prep-stable list when cycle time is tight. Discovery detectability does not prove targeted response after cleanup (Carr et al. 2014). Absolute units still reflect the surrogate under the stated spike timing.
What Sample-Matched Targeted Analysis Can Support
Matched prep and a locked relative method support same-peptide comparison across contrast arms. Matched prep plus SIS design supports absolute amount for surrogates inside the calibrated interval.
Failed targets after a matrix pilot support peptide redesign, cleanup revision, or a PRM subset. They do not support silent zeros. Prep notes belong with the quantitative table so batch structure is reviewable. Cross-study comparison needs matching prep and normalization or calibration assumptions.
A sample-matched targeted package supports the declared panel decision. It does not rescue biology from unmatched pretreatment.
How to Choose Sample and Prep Scope
Use matrix risk, prep parity, panel status, and claim unit together.
Sample and prep signal |
Scope that often fits |
Recheck before committing |
|---|---|---|
Matched lysate prep, small closed panel |
MRM or PRM after matrix-matched lock |
Ship full cohort before digest pilot |
Plasma or tissue with new cleanup |
Staged prep plus method pilot |
Assume lysate methods transfer |
Depletion or pull-down used |
Same enrichment on every arm |
Enrichment on treated arm only |
Study n large, QC omitted from count |
Recalculate injection load |
Quote on study n alone |
Amount claim after digest SIS |
Peptide-level calibration only |
Treat as protein extract amount |
Prep-stable MRM, few peptides fail |
Hybrid MRM plus PRM subset |
Whole-panel PRM without need |
Open protein list, prep still moving |
Discovery first; freeze prep before panel |
Targeted quote on unnamed targets |
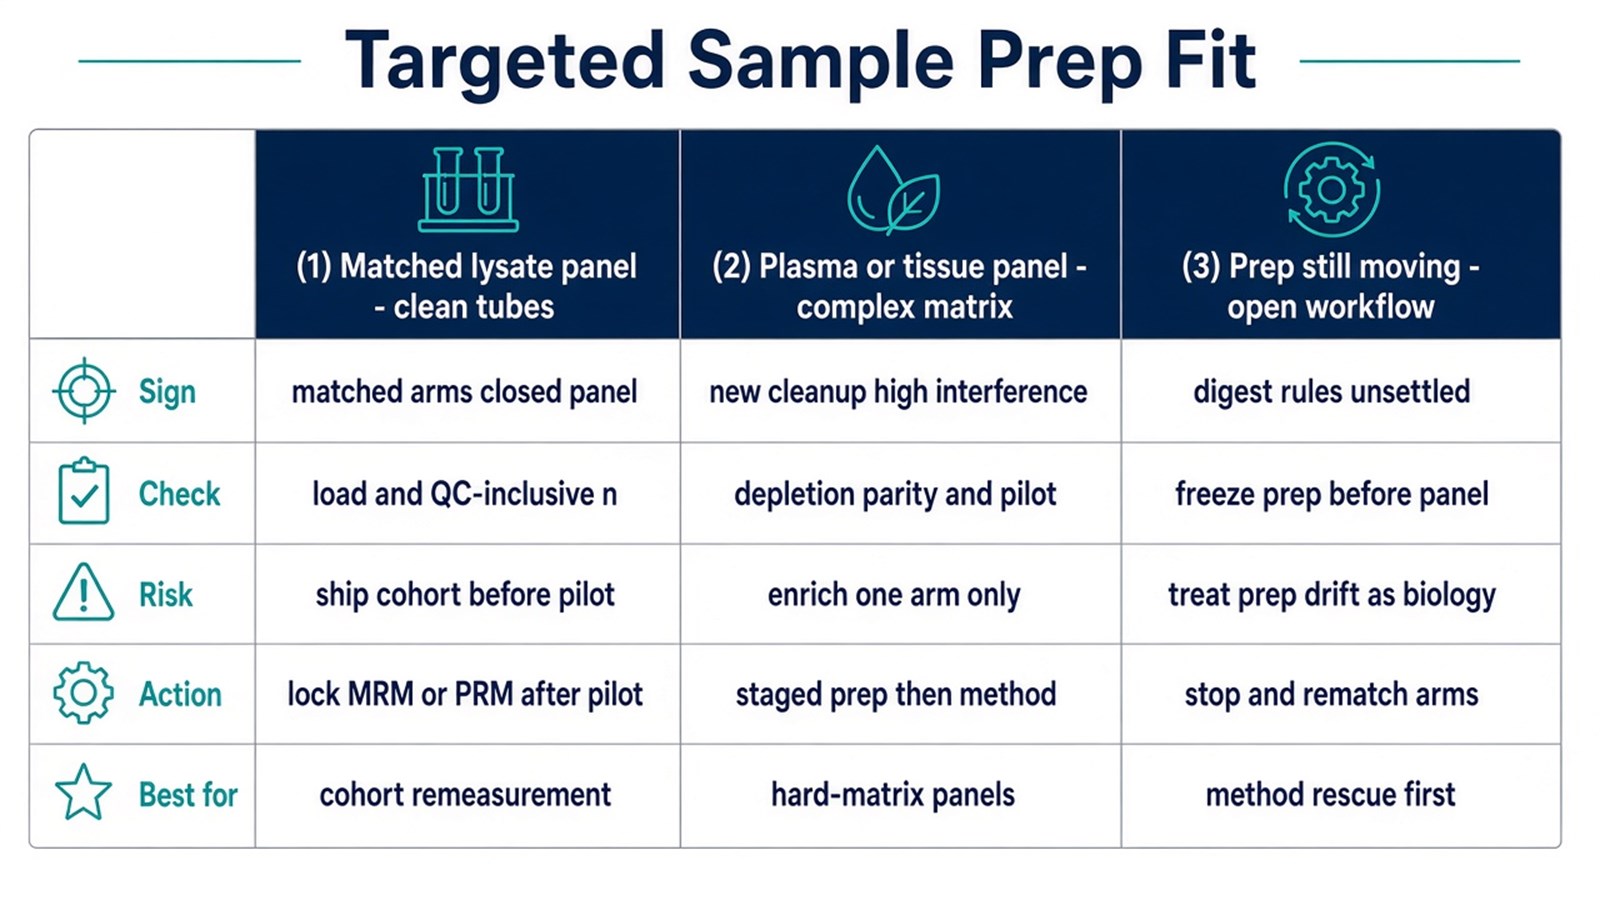
Figure 2. Match targeted analysis scope to pretreatment parity and a matrix-matched pilot; keep QC and SIS injections inside the sample count.
Quote-stage panel and deliverable lock appears in Scoping a Targeted Mass Spectrometry Project: Analyte Panel, Platform, and Validation Deliverables. Teams with matrix documentation, pretreatment records, closed peptide tiers, and QC-inclusive sample counts can review MtoZ Biolabs Targeted Proteomics Service when those fields are declared in the project record.
Frequently Asked Questions
1. Why does targeted proteomics analysis begin with sample and prep?
Because scheduled transitions or fragment rules only hold on peptides that survive a declared matrix and pretreatment route. Unmatched digest or cleanup can create false biological contrast.
2. How many samples are needed for a targeted study?
There is no fixed n. Count biological replicates required by the contrast, then add blanks, QC pools, and any SIS or calibration levels. Study n alone understates injection load.
3. Can methods be optimized on neat peptide standards only?
Neat standards help choose starting conditions. Matrix-matched digests after the declared prep are required before cohort lock.
4. Does a larger panel reduce the need for matched pretreatment?
No. A larger panel increases cycle-time pressure. Unmatched digest or cleanup still shifts peptide pools across arms.
5. When should prep problems move peptides from MRM to PRM?
When matrix-matched pilots still show unstable transition ratios after cleanup and collision-energy work, a PRM subset can be scoped. Prep mismatch should be fixed first.
MtoZ Biolabs can provide targeted proteomics analysis using MRM, PRM, and isotope-assisted absolute quantification for predefined protein or peptide panels, including matrix-matched sample preparation and QC-supported reporting when panel design and study goals are documented.
If further evaluation of digest pilots, enrichment parity, SIS spike timing, or hybrid MRM plus PRM panels is needed, project details can be submitted for a feasibility review.
Reference
- V. Lange, P. Picotti, B. Domon, R. Aebersold (2008). Selected reaction monitoring for quantitative proteomics: a tutorial. Mol. Syst. Biol., 4, 222. https://doi.org/10.1038/msb.2008.61
- A.C. Peterson, J.D. Russell, A.A. Klammer, P.F. Doubleday, H. Liu, J.N. Anderson, M.J. MacCoss (2012). Parallel reaction monitoring for high resolution and high mass accuracy quantitative, targeted proteomics. Mol. Cell. Proteomics, 11, 1475-1488. https://doi.org/10.1074/mcp.O112.020271
- S.A. Gerber, J. Rush, O. Stemman, M.W. Kirschner, S.P. Gygi (2003). Absolute quantification of proteins and phosphoproteins from cell lysates by tandem MS. Proc. Natl. Acad. Sci. U.S.A., 100, 6940-6945. https://doi.org/10.1073/pnas.0832254100
- S.A. Carr, S.E. Abbatiello, B.L. Ackermann, C. Borchers, B. Domon, E.W. Deutsch, R.P. Grant, A.N. Hoofnagle, R. Huttenhain, J.M. Koomen, D.C. Liebler, T. Liu, B. MacLean, D.R. Mani, E. Mansfield, H. Neubert, A.G. Paulovich, L. Reiter, O. Vitek, R. Aebersold, A.J. Smith, et al. (2014). Targeted peptide measurements in biology and medicine: best practices for sample preparation, assay standardization, and assay validation. Mol. Cell. Proteomics, 13, 907-917. https://doi.org/10.1074/mcp.M113.036095
- M. Bantscheff, S. Lemeer, H.U. Haab, B. Kuster (2012). Quantitative mass spectrometry in proteomics: a critical review. Anal. Bioanal. Chem., 404, 939-965. https://doi.org/10.1007/s00216-012-5892-4
