



A tandem mass tag (TMT) is an isobaric chemical label that marks peptides from different samples so they can be mixed and compared in one liquid chromatography-tandem mass spectrometry (LC-MS/MS) run (Thompson et al. 2003). Each tag in a kit has the same nominal mass at the precursor level. Fragmentation releases a reporter ion whose mass identifies the channel, and reporter intensities become relative peptide, then protein, values. Tandem mass tag quantitative proteomics is that labeling experiment. It is not a colorimetric total-protein assay, not an intact-mass identity check, and not an absolute concentration method. Search titles such as Tandem Mass Tag (TMT)-Based Quantitative Proteomics name the same reagent class. They do not name a different acquisition mode.
Researchers who already have defined samples, comparison groups, or a specific project objective can also review the MtoZ Biolabs TMT Quantitative Proteomics Analysis Service for project-specific feasibility and analysis planning.
What a Tandem Mass Tag Is
A commercial TMT reagent has three parts: an amine-reactive group, usually an N-hydroxysuccinimide (NHS) ester, a mass-normalizing balancer, and a reporter. The reactive group labels peptide N-termini and lysine residues when the digest is compatible with that chemistry. The balancer offsets the reporter mass so that every channel stays isobaric until fragmentation. After mixing, the same peptide from all channels co-elutes as one precursor. MS2, or MS3 on instruments that support it, records reporter ions in the low-mass region of the spectrum.
Kit formats change the number of channels that can share that precursor. Commercially available TMT reagents have been supplied as 6-plex, 10-plex, 11-plex, and TMTpro 16-plex or 18-plex sets, depending on the reagent generation. The useful set is the one that holds the planned groups and replicates without forcing unrelated samples into leftover channels. An 18-channel kit does not make the method an open cohort assay. Mixing still closes the analytical block.
Some TMTpro reporter pairs are only a few millidaltons apart. Those channels need sufficient resolving power to keep the reporters distinct. Millidalton spacing is a hardware requirement for that kit, not proof that every TMT ratio is more accurate than an unlabeled peak area.
TMT is a product name for this isobaric class. The scientific operation is amine-directed isobaric tagging, pooling, and reporter-ion comparison.
Researchers who need to determine whether that operation matches a study can refer to TMT-Based Proteomics for more detailed guidance.
A tandem mass tag codes a closed channel set into one precursor mass; it does not create an absolute protein amount.
How Tag Structure Becomes a Protein Ratio
After extraction and trypsin digestion, each sample receives one channel. Unused tag is removed or quenched so that leftover reagent does not label the wrong digest after mixing. Labeled peptides are combined, often fractionated when the mixture is dense, and acquired by LC-MS/MS. Sequence ions identify the peptide. Reporter ions quantify the channels. Protein values are inferred from those peptides, so shared peptides, missed cleavage, and incomplete labeling can move the reported number.
Acquisition is a separate decision. In MtoZ isobaric-labeling offerings, TMT workflows are typically acquired in data-dependent acquisition (DDA) so reporter ions can be read. Data-independent acquisition (DIA) is not a TMT step and is not a required partner method. A labeling efficiency percentage is a process check for that batch. It is not a universal property of the reagent.
Researchers who need those stage-level controls can refer to A Detailed Workflow of TMT-Based Quantitative Proteomics for load matching, fractionation, and reporter readout.
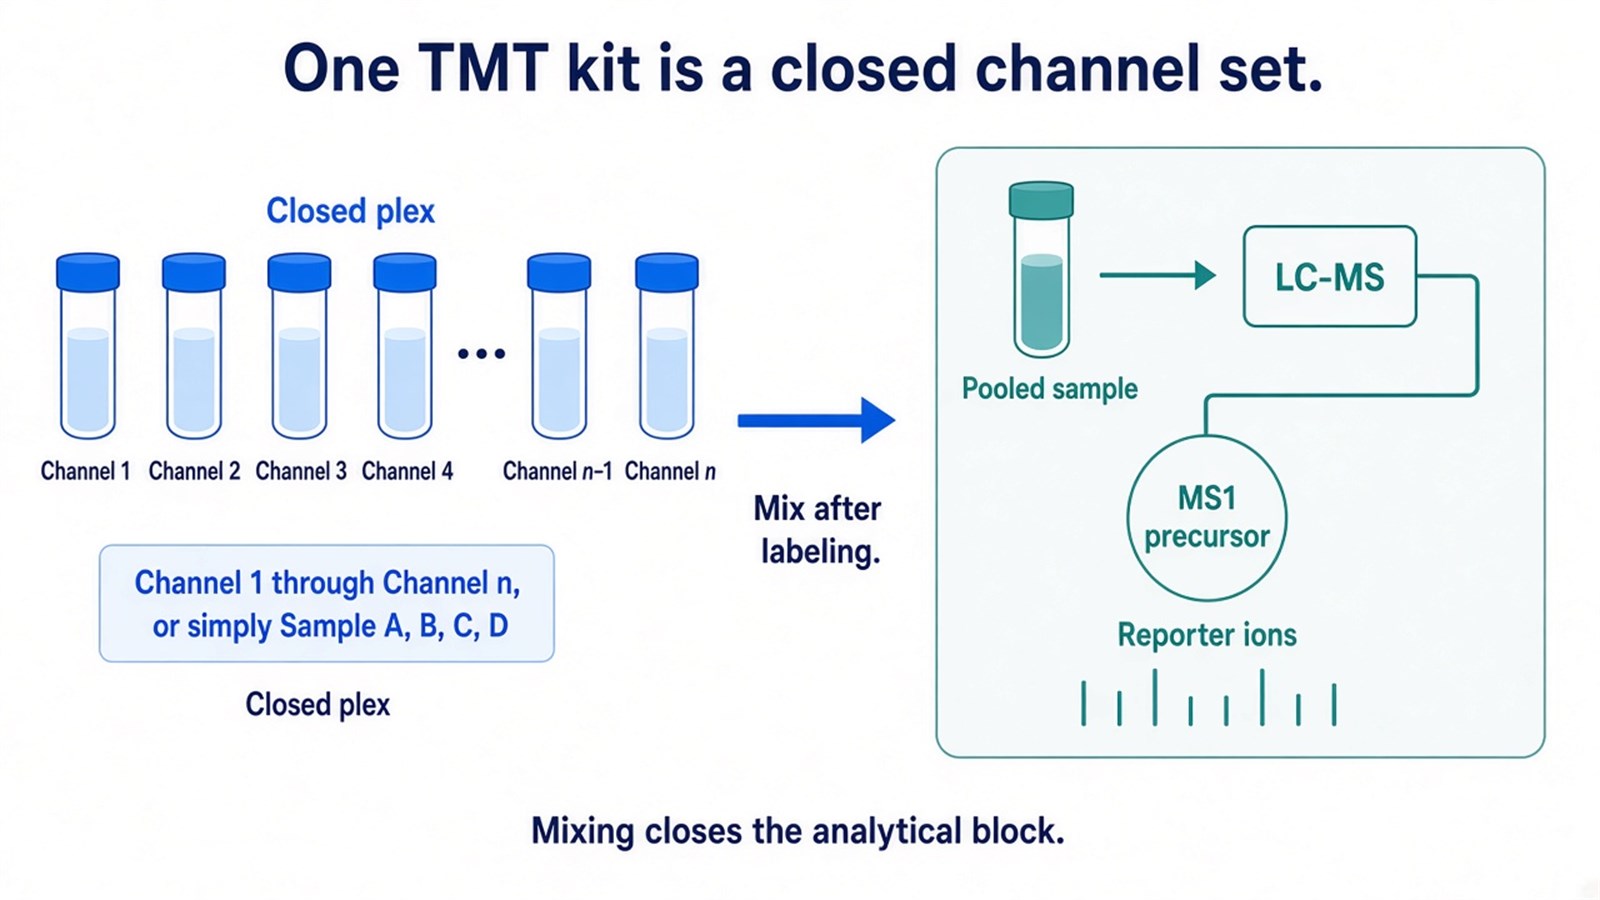
Figure 1. One TMT kit is a closed channel set; labeled peptides share one MS1 mass, and reporter ions carry the relative comparison after fragmentation.
Plex Size, Bridge Channels, and What the Tag Cannot Stretch
A single plex fits a contrast that can be frozen before labeling. Three groups with biological replicates can sit in one 10-plex or 16-plex when leftover channels are not filled with unrelated samples. Randomizing group labels across channels avoids putting every control on adjacent reporters.
A cohort larger than one kit becomes several plexes. Each new kit is a new analytical block. A pooled bridge sample, labeled in every plex, can help estimate inter-plex scale. The bridge does not turn two instruments, two digest batches, or two enrollment waves into one experiment. Ordinary reporter normalization does not insert a sample that arrived after mixing into the original matrix.
Unlabeled label-free quantification remains the better match when samples must stay as independent injections or when enrollment will continue. Saying that TMT reduces batch effects is true only inside one mixed run. It is false as a claim that TMT is the large-cohort method and LFQ is the small-cohort method.
Tag decision |
What it fixes |
What it does not fix |
|---|---|---|
Choose a kit that holds the contrast |
Channel count for one closed set |
Later samples after mixing |
Randomize groups across channels |
Channel-position bias |
Incomplete labeling |
Add a bridge channel across plexes |
A scale reference between kits |
A new digest or instrument block |
Use TMTpro for more channels |
A larger closed set |
An open, growing cohort |
High resolving power for close reporters |
Separation of millidalton reporter pairs |
Ratio compression from co-isolation |
Co-isolation can still compress fold changes toward one (Ting et al. 2011). SPS-MS3 can reduce that mixing on instruments that support the scan (McAlister et al. 2014). Those limits come from isolation and fragmentation, not from the printed plex number on the box.
What Tandem Mass Tag Quantification Can Support
After identification, reporter extraction, impurity correction, and protein summarization, a TMT value can support a relative statement: protein A is higher or lower in channel 1 than in channel 2 inside this kit. Peptide support and the stated plex map belong next to that statement. Ordinary TMT ratios should not rank different proteins by copy number, infer a diagnostic cutoff, or transfer a fold change from one kit generation to another study as a calibrated quantity.
Amine-reactive tags can be used on peptides after a modification enrichment, for example a phosphopeptide enrichment. The tag still reports relative channel intensity of the enriched peptide. It does not, by itself, map a signaling network or prove target engagement. Tissue, cell, biofluid, or immunoprecipitate matrices can be labeled when amines are available and buffers do not quench the NHS ester. Naming those matrices does not change the relative-only claim.
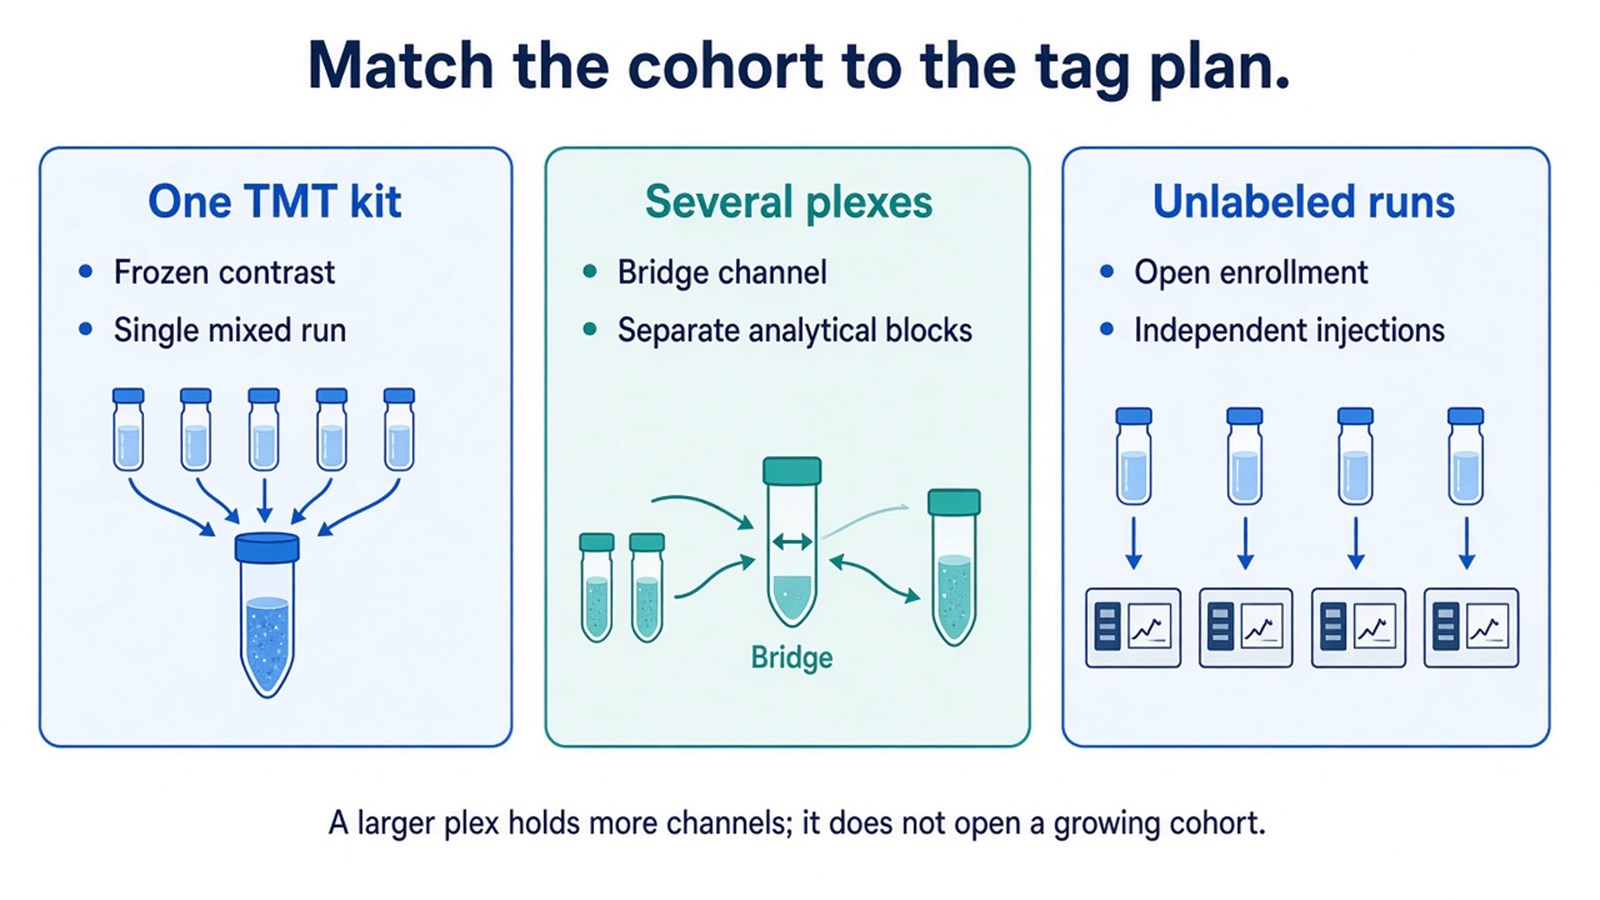
Figure 2. One kit holds a frozen contrast; a bridge links separate plexes without erasing the block; an open enrollment stays unlabeled.
Tandem mass tag quantitative proteomics supports relative ratios inside a closed plex map, not a growing unlabeled series.
Frequently Asked Questions
1. Is a tandem mass tag the same as TMT-based proteomics?
The tag is the reagent. TMT-based proteomics is the labeling, mixing, and reporter-ion experiment that uses that reagent. The two names are usually used for the same relative multiplex study.
2. Does a higher plex number mean better quantification?
No. A larger kit holds more channels in one mixed run. It does not reduce co-isolation, and it does not license leftover channels filled with unrelated samples.
3. Can TMT quantitative proteomics report absolute protein amounts?
Not from ordinary reporter-ion tables. Amount claims need a matched standard and a calibration model, usually in a targeted experiment.
4. If more samples arrive after the first 16-plex, can they join the same TMT table?
Not as additional channels in the original mix. They need a new kit. A bridge sample can help compare plexes. It does not rewrite the first reporter matrix.
Once the kit map, labeling chemistry, and expected protein-level readout are defined, the project can be evaluated as a tandem mass tag study. Researchers can review the MtoZ Biolabs TMT Quantitative Proteomics Analysis Service for project-specific sample evaluation, feasibility assessment, and workflow planning.
