



Specific protein quantification means measuring named proteins under a declared claim unit—not surveying an open proteome. In mass spectrometry, that usually means targeted proteomics: proteotypic peptides scheduled by selected reaction monitoring (SRM), multiple reaction monitoring (MRM), or parallel reaction monitoring (PRM) on liquid chromatography-tandem mass spectrometry (LC-MS/MS) (Lange et al. 2008; Peterson et al. 2012). Enzyme-linked immunosorbent assay (ELISA) and Western blot quantify antibody-defined epitopes and answer a different design object. In project language, “specific” starts with a closed protein or peptide list, matched sample prep, and a relative versus absolute decision. The guide below maps comprehensive targeted-MS framing for specific protein quantification with workflow, locked parameters, limits, and fit versus antibody methods or discovery. It is not a mid-century methods history, not a clinical diagnostics catalog, and not a claim that any MS run is automatically absolute.
Relative fold change and amount with stable isotope-labeled internal standard (SIS) peptides remain separate claim units (Gerber et al. 2003). Unmatched prep still breaks specificity after peak integration (Bantscheff et al. 2012).
In MtoZ Biolabs targeted offerings, specific protein quantification is scoped after target list, matrix, claim unit, platform preference, and quality control (QC) design are written in the brief. Service intake appears in Targeted Proteomics Service.
What Specific Protein Quantification Must Cover
Four layers belong in the brief and stay separated.
Target-intake layer freezes protein names and the proteotypic peptides or epitopes used as surrogates. Open discovery breadth is outside this specific record.
Claim layer names relative panel tracking versus SIS absolute concentration. Discovery intensities and blot band darkness are not automatic amount units.
Method-class layer names targeted MS (MRM/PRM), ELISA, or Western blot. Peptide surrogates and antibody epitopes are not interchangeable evidence.
Prep-and-QC layer names matched extraction or lysis, digest when MS is used, blanks, and failed-target or calibration rules.
Specific quantification remonitors declared analytes. It does not invent unnamed proteins or convert every signal into concentration.
Trade-offs of the targeted-MS class appear in Advantages and Limitations of Targeted Proteomics. Absolute calibrator scoping appears in Absolute Protein Quantification.
How Targeted MS Compares with ELISA and Discovery
Targeted MS, ELISA, and discovery answer different specificity objects.
MRM/PRM measure scheduled peptides with assay-level QC after matrix pilots. They fit when antibodies are unavailable, isoform or peptide context matters, or multiplex closed panels must be remonitored across a cohort. PRM versus MRM choice appears in Parallel Reaction Monitoring vs MRM: Which Fits?.
ELISA fits when a validated antibody pair already answers the epitope claim at the needed throughput. Western blot fits low-plex confirmation of size/context, not large cohort absolute tables by default.
Discovery LFQ, DIA, or labeled discovery still comes first when the protein list is open. When targeted remonitoring should replace rediscovery appears in When Targeted Mass Spectrometry Outperforms Discovery.
End-to-End Workflow for Specific Protein Quantification by Targeted MS
A practical targeted spine for named proteins usually runs as follows.
- Freeze the named proteins, contrast, and relative versus absolute claim.
- Map proteotypic peptides and uniqueness rules after the declared digest.
- Declare matrix and matched prep for every contrast arm.
- Choose MRM, PRM, or a mixed panel from interference evidence.
- Build transitions or precursor windows on matrix-matched digests; add SIS and calibrators only if amount is required.
- Set dwell times or retention windows against peak width and cycle time.
- Size biological n plus blanks, QC pools, and calibration injections if used.
- Acquire interleaved study and QC injections; stop if ratios, retention, or missingness break locked rules.
- Report only the predefined proteins under the declared claim unit; archive method and list versions.
Steps 1 through 4 protect specificity. Steps 5 through 9 produce the quantitative record.
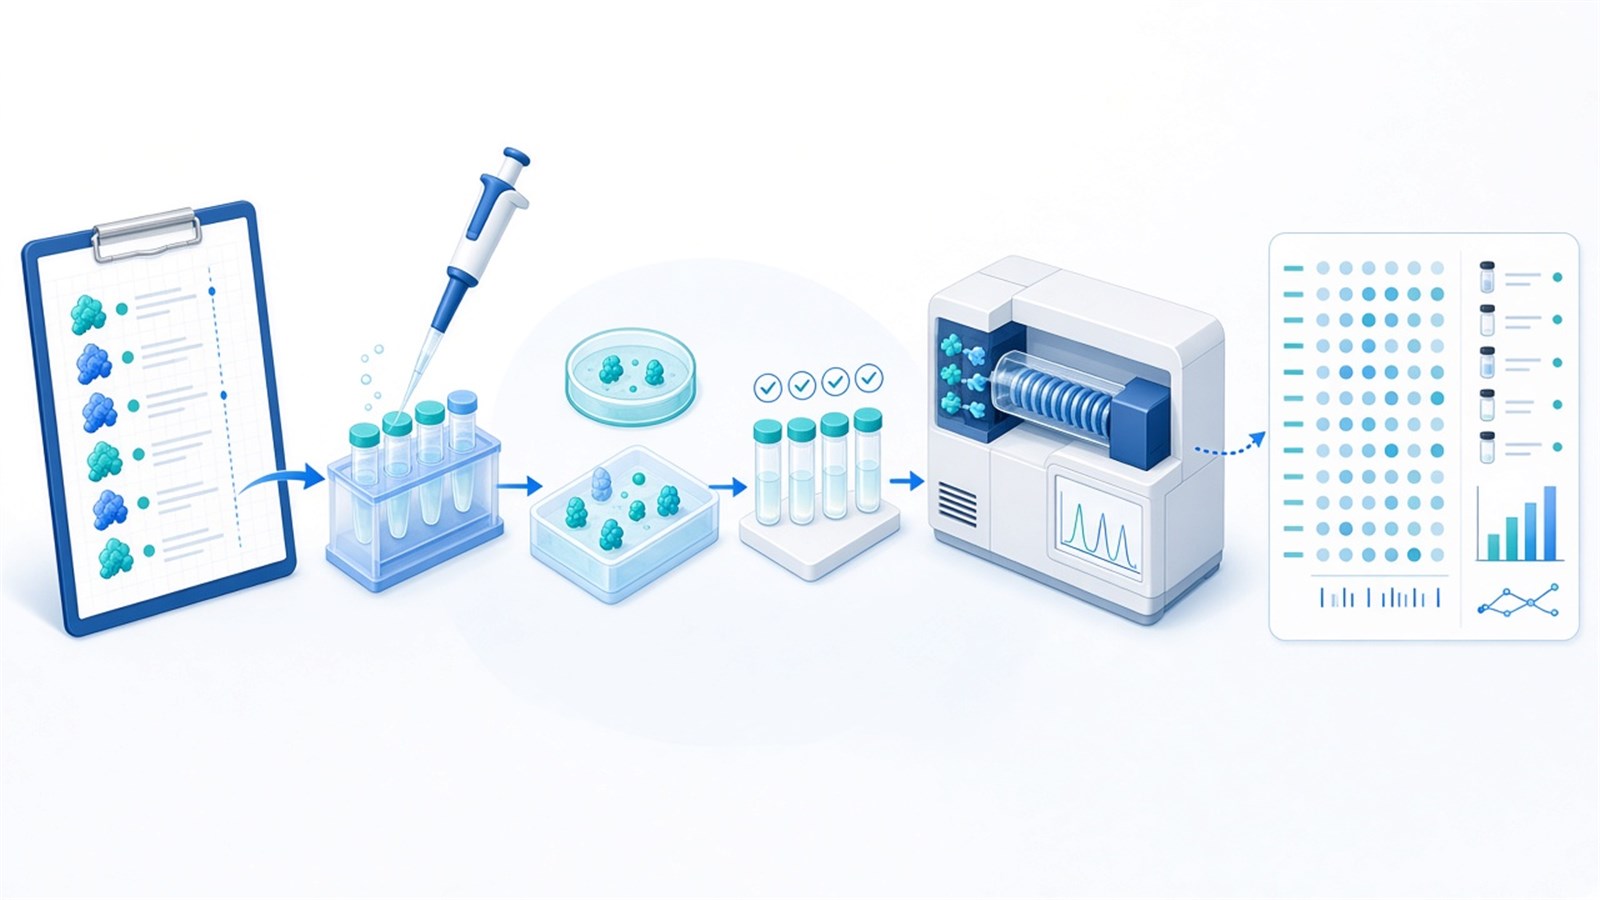
Figure 1. Specific protein quantification by targeted MS depends on a closed list and matched prep before contrasts are tested.
Parameters That Must Be Locked
Layer |
Parameter that must be declared |
Failure mode if ignored |
|---|---|---|
Targets |
Protein and peptide list version |
Moving “specific” set mid cohort |
Claim unit |
Relative vs SIS absolute |
Band or peak height as concentration |
Matrix |
Sample type, harvest, storage |
Suppression as biology |
Prep |
Digest or lysis recipe parity |
Unequal recovery across arms |
Platform |
MRM, PRM, ELISA, or blot rationale |
Wrong specificity object |
Schedule |
Cycle time vs peak width (MS) |
Undersampled transitions |
SIS |
Spike step when absolute |
Relative peaks sold as amount |
QC |
Pools, blanks, failed-target rules |
Undetected drift |
Antibody (if used) |
Clone/epitope documentation |
Epitope ≠ peptide claim |
Deliverable |
Method and list archive |
Report rebuilt after acquisition |
Limits follow the same table. MS high-throughput slogans do not remove cycle-time ceilings. ELISA specificity is epitope-bound. Western blot is not a default absolute assay. Clinical diagnosis language needs separate study design beyond the specific-protein table.
How to Choose a Route for Specific Proteins
Use list status, reagent constraints, claim unit, and cohort need together.
Project need |
Route that often fits |
Recheck before committing |
|---|---|---|
Closed proteins, no suitable antibody, cohort remonitoring |
Targeted MRM/PRM |
Force ELISA without epitope reagents |
Validated antibody already answers the claim |
ELISA |
Rebuild MS for the same epitope only |
Open proteome candidate finding |
Discovery LFQ/DIA first |
“Specific” MS on unnamed targets |
Amount on named peptides |
Targeted MS + SIS calibrators |
Relative MS or blot as concentration |
Interference on MRM channels |
PRM subset or peptide redesign |
Keep failing transitions |
Low-plex size/context check |
Western blot support |
Large absolute cohort by blot alone |
Prep recipe still changing |
Stop method lock |
Acquisition during prep drift |
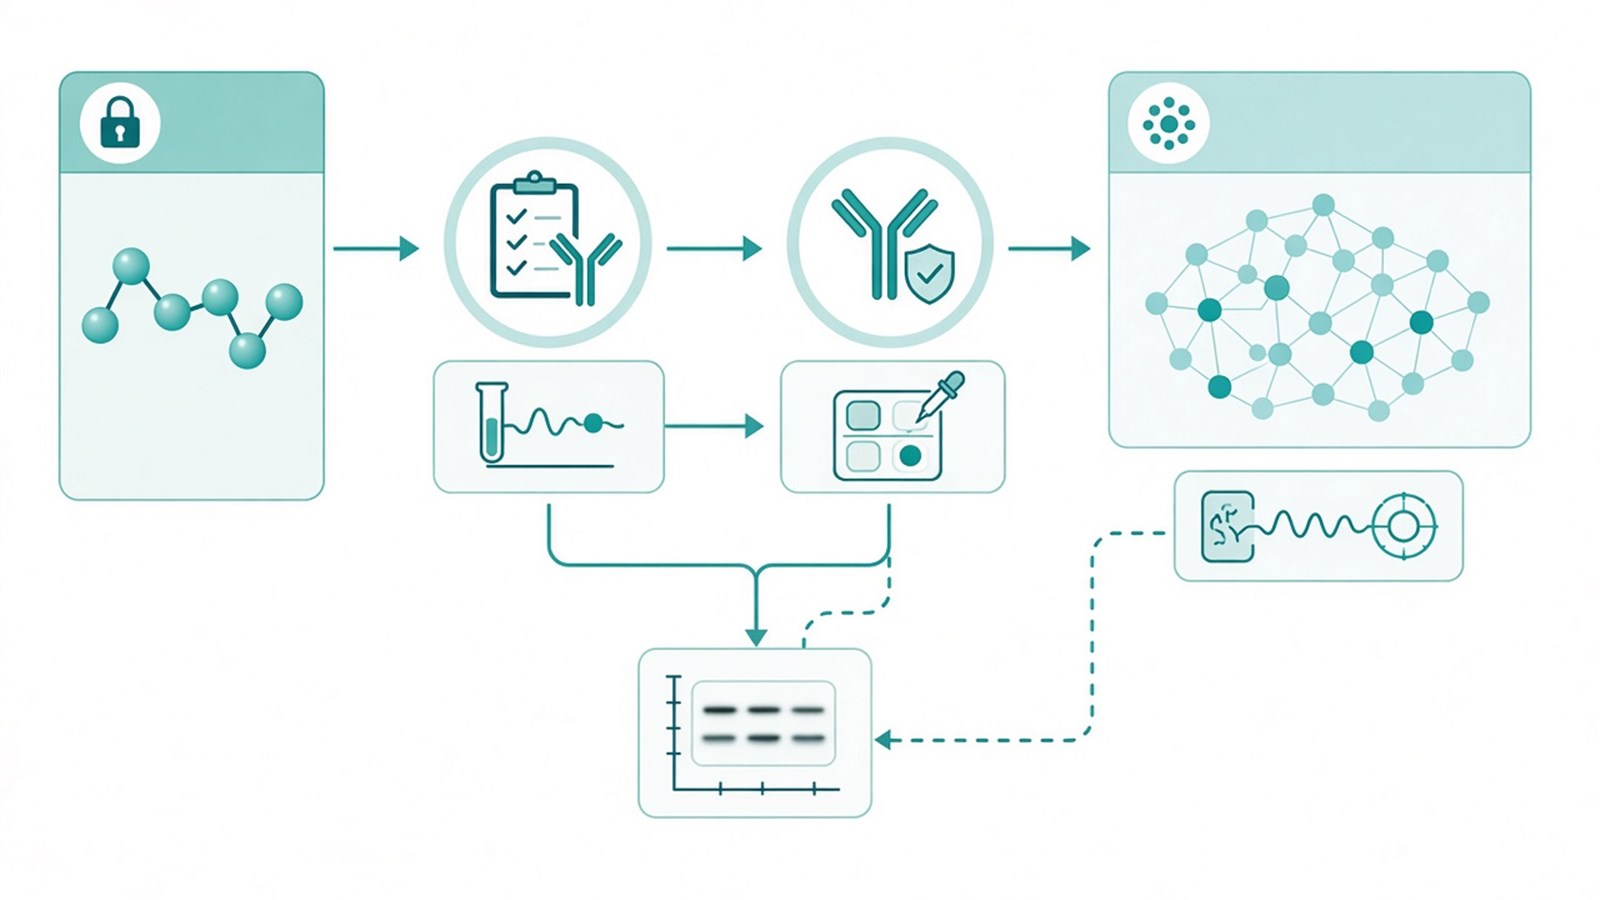
Figure 2. Choose targeted MS for closed peptide surrogates; keep ELISA when epitopes already answer the claim; keep discovery while the list is open.
Teams with named protein lists, matrix notes, and claim units can review MtoZ Biolabs targeted and MRM service entries when those fields are declared in the project record.
Frequently Asked Questions
1. Is specific protein quantification the same as discovery proteomics?
No. Specific work starts from named analytes. Discovery finds candidates before that list is closed.
2. Does mass spectrometry always beat ELISA for specific proteins?
No. ELISA wins when epitope reagents already answer the claim. Targeted MS wins when peptide surrogates, multiplex panels, or missing antibodies fit better.
3. Can Western blot provide absolute concentration?
Not by default. Blot signals need separate calibration designs and still report epitope-linked evidence.
4. Must every specific MS assay use SIS peptides?
No. Relative contrasts can run light-only when that claim is declared. SIS and curves become necessary for amount units.
5. When should projects stay in discovery instead?
When the protein list is still open or proteotypic surrogates are not yet proven in matrix.
MtoZ Biolabs can discuss specific protein quantification scoping when target list, matrix, and claim unit are documented.
If further evaluation of MRM versus PRM, SIS absolute design, or ELISA fit is needed, project details can be submitted for a feasibility review.
Reference
- V. Lange, P. Picotti, B. Domon, R. Aebersold (2008). Selected reaction monitoring for quantitative proteomics: a tutorial. Mol. Syst. Biol., 4, 222. https://doi.org/10.1038/msb.2008.61
- A.C. Peterson, J.D. Russell, A.A. Klammer, P.F. Doubleday, H. Liu, J.N. Anderson, M.J. MacCoss (2012). Parallel reaction monitoring for high resolution and high mass accuracy quantitative, targeted proteomics. Mol. Cell. Proteomics, 11, 1475-1488. https://doi.org/10.1074/mcp.O112.020271
- S.A. Gerber, J. Rush, O. Stemman, M.W. Kirschner, S.P. Gygi (2003). Absolute quantification of proteins and phosphoproteins from cell lysates by tandem MS. Proc. Natl. Acad. Sci. U.S.A., 100, 6940-6945. https://doi.org/10.1073/pnas.0832254100
- M. Bantscheff, S. Lemeer, H.U. Haab, B. Kuster (2012). Quantitative mass spectrometry in proteomics: a critical review. Anal. Bioanal. Chem., 404, 939-965. https://doi.org/10.1007/s00216-012-5892-4
