Dimethyl labeling Service
MtoZ Biolabs provides Dimethyl labeling Service for relative protein quantification by stable-isotope peptide labeling and LC-MS/MS analysis across defined experimental groups.
Light, intermediate, and heavy dimethyl labels encode digested peptides before sample mixing, enabling MS1-based abundance comparison with peptide and protein identification.
- Stable-isotope dimethyl labeling at the peptide level
- Light, intermediate, and heavy labeling options
- Protein identification and differential analysis
What Is Dimethyl Labeling?
Dimethyl labeling is a relative quantitative proteomics strategy based on stable-isotope chemical labeling. After protein digestion, peptides from different samples are separately dimethylated using formaldehyde and reducing reagents with different isotopic compositions, generating mass differences that allow mass spectrometry to distinguish peptides with the same sequence. After labeling, the different samples are mixed according to the study design and analyzed by LC-MS/MS. Relative abundance is calculated from the MS1 signal intensity or peak-area ratios of the corresponding isotopically labeled peptides.
Unlike SILAC, which depends on metabolic incorporation in living cells, Dimethyl labeling performs chemical labeling at the peptide level and therefore does not require sample sources that are compatible with metabolic labeling. Classical designs can use duplex or triplex isotopic labels for relative comparison of a small number of clearly defined sample groups within the same analytical batch. The primary purpose of Dimethyl labeling is to compare relative protein abundance rather than directly determine absolute protein concentrations.
When This Service Is a Good Fit
Dimethyl labeling can be considered for the following study designs:
- Global relative protein abundance needs to be compared among a small number of clearly defined experimental conditions or treatment groups.
- Stable-isotope labeling is required, and differently labeled samples are mixed after labeling for analysis within the same LC-MS/MS run.
- The sample source is not suitable for SILAC metabolic labeling, but a mass-difference-based stable-isotope quantification strategy is still required.
- The study requires protein identification, relative quantification, and differential protein results, followed by candidate protein screening.
- A PTM proteomics study or another enrichment-based study already has an established sample-preparation workflow and requires an additional stable-isotope relative quantification step at the peptide level.
For studies with sample numbers that substantially exceed the capacity of classical duplex or triplex labeling, the suitability of batch-based Dimethyl labeling needs to be evaluated according to the batch design and quantitative objectives. Large sample cohorts generally also require comparison with DIA, TMT, or other quantitative strategies.
Technical Principles
1. Reductive Dimethylation of Peptides
Dimethyl labeling is based on reductive amination. Formaldehyde reacts with the α-amino group at the peptide N-terminus and the ε-amino group on lysine side chains, forming stable dimethylated products in the presence of a reducing reagent. Because most tryptic peptides contain free amino groups available for labeling, reductive dimethylation enables broad isotopic encoding at the peptide level.
2. Stable-Isotope Multiplexing
Formaldehyde and reducing reagents with different isotopic compositions can be selected to generate light, intermediate, and heavy mass labels. Peptides with the same sequence from different samples have similar chemical properties and chromatographic behavior but appear as distinguishable mass peaks in MS1, allowing simultaneous identification and comparison within a mixed sample.
3. MS1-Based Relative Quantification
After labeled samples are mixed, LC-MS/MS analysis is performed. Protein identification relies on peptide MS/MS evidence, while relative quantification is primarily calculated from the MS1 peak areas or signal intensities of the corresponding light, intermediate, and heavy peptide ions. Quantitative information from multiple peptides is then summarized at the protein level for between-group relative abundance comparisons.
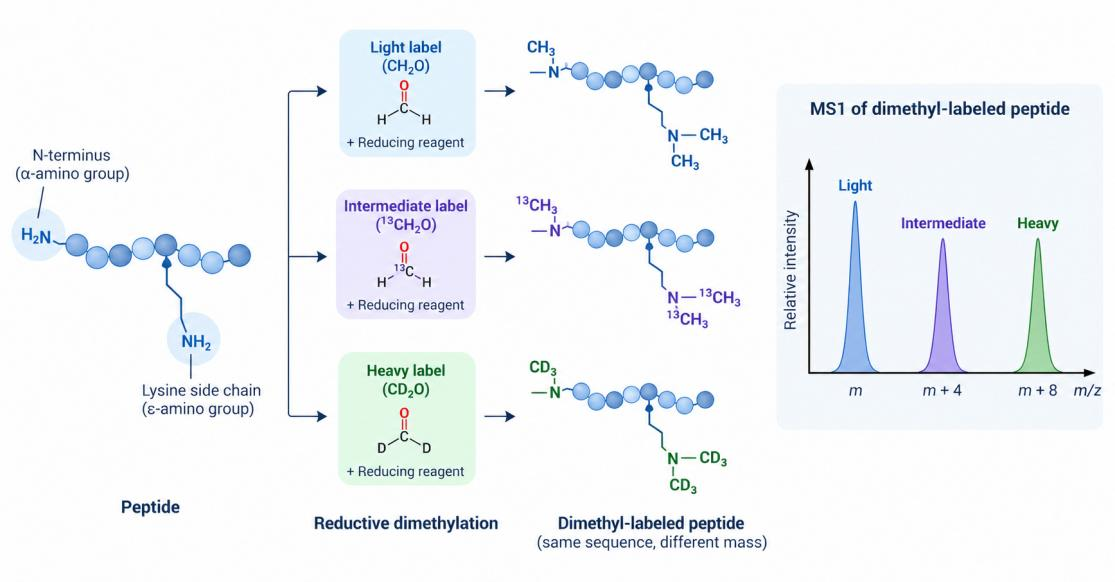
Figure 1. Stable Isotope Dimethyl Labeling Principle.
Dimethyl Labeling Service at MtoZ Biolabs
MtoZ Biolabs provides Dimethyl labeling Service with protein extraction, protein quantification, digestion, stable-isotope dimethyl labeling of peptides, labeled-sample mixing, LC-MS/MS acquisition, and quantitative data processing arranged according to study groups and sample conditions. Analysis results can include peptide and protein identification, relative abundance, differential proteins between groups, and quality-control information. Functional annotation or other extended analyses are determined according to the project analysis scope. Before a project begins, sample number, sample matrix, available protein amount, buffer system, and research objectives can be evaluated to determine a suitable labeling and comparison design.
Analysis Workflow
1. Sample and Study Design Evaluation
Confirm the sample type, group design, sample number, available sample amount or protein amount, and primary quantitative objectives.
2. Protein Extraction and Quantification
Perform protein extraction and concentration evaluation according to sample condition to establish a basis for subsequent digestion and equal-amount labeling.
3. Protein Digestion and Peptide Preparation
Digest proteins into peptides and perform necessary desalting or purification to obtain peptide samples suitable for chemical labeling.
4. Dimethyl Labeling
Label peptides from different samples separately with the corresponding stable-isotope dimethyl labels and perform labeling quality checks.
5. Sample Mixing and LC-MS/MS
Mix differently labeled samples according to the experimental design, separate the peptides by liquid chromatography, and perform MS and MS/MS acquisition.
6. Protein Identification and Relative Quantification
Complete peptide and protein identification, calculate relative abundance from isotopically labeled peptide signals, and perform differential analysis and result organization according to the study groups.
Sample Submission Suggestions
Key sample considerations for Dimethyl labeling include not only the available protein amount but also sample-processing status, buffer composition, and additive compatibility. During project consultation, researchers are recommended to provide the sample type, group design, sample number, available sample amount or protein amount, protein extraction or digestion status, buffer composition, and research objectives so that the labeling design and LC-MS/MS analysis conditions can be evaluated.
|
Sample Status |
Suggested Information |
|
Tissue/Cell/Microbial Samples |
Species or source, sample type, group design, biological replicates, storage conditions, available sample amount, and primary research objectives. |
|
Serum/Plasma/Other Biofluids |
Sample type, collection and storage conditions, freeze-thaw history, group design, available volume, and whether high-abundance protein processing has been performed. |
|
Protein Extracts |
Extraction method, buffer system, protein concentration/total amount, information on detergents or other additives, storage conditions, and group design. |
|
Digested Peptides |
Digestion method, total peptide amount, desalting/purification status, current solution composition, group design, and whether any additional enrichment has been performed. |
Dimethyl labeling depends on chemical reactions involving peptide amino groups. Complete information on the buffer system and additives should therefore be provided for extracted proteins or digested peptides so that components that may affect the labeling reaction can be evaluated.
Why Choose MtoZ Biolabs?
1. Customized Service
MtoZ Biolabs evaluates suitable Dimethyl labeling designs according to sample number, group structure, sample condition, and quantitative objectives, with corresponding labeling and between-group comparison strategies selected for the study.
2. One-Stop Analysis
MtoZ Biolabs integrates sample evaluation, protein processing, and digestion with Dimethyl labeling, sample mixing, LC-MS/MS acquisition, and quantitative analysis, reducing coordination requirements across different experimental stages.
3. High-Resolution LC-MS/MS
High-resolution LC-MS/MS provides peptide identification and MS1 quantitative signals corresponding to different isotopic labels, establishing the data basis for relative protein abundance comparisons.
4. Proteomics Expertise
Experience in quantitative proteomics and mass spectrometry supports comprehensive evaluation of sample condition, labeling design, mass spectrometry acquisition, and result quality, providing technical support for different study designs.
Applications
1. Disease-Related Proteomics Research
Compare relative protein abundance among different disease models, biological states, or research groups to provide quantitative data for disease-related proteomics research.
2. Drug and Treatment Research
Compare protein abundance changes following drug treatment, stimulation, intervention, or other treatment conditions to support research on drug effects and treatment responses.
3. Cellular Regulation Research
Analyze protein expression changes under different cellular states, genetic perturbations, stress conditions, or culture conditions for studies of cellular regulation and biological processes.
4. PTM Quantitative Proteomics
When the corresponding modified-peptide enrichment and experimental design are suitable, Dimethyl labeling can be used for between-group relative quantification in phosphoproteomics, acetylproteomics, and other PTM proteomics studies.
5. Plant and Microbial Proteomics
Dimethyl labeling can be used for comparative proteomics of plant, microbial, and other research samples to investigate protein abundance changes associated with different growth, environmental, or treatment conditions.
Deliverables
1. Experimental Workflow, Dimethyl Labeling Design, and LC-MS/MS Analysis Information
2. Peptide and Protein Identification Results
3. Dimethyl-Labeled Peptide Signals and Relative Protein Quantification Results
4. Between-Group Differential Protein Statistics and Quality-Control Information
5. Functional Annotation or Other Extended Analysis Results (If Applicable)
6. Raw Mass Spectrometry Data, Processed Data Files, and Analysis Report
FAQ
Q1: Is Dimethyl labeling performed at the protein level or peptide level?
In conventional quantitative proteomics, Dimethyl labeling is typically performed on peptides after protein digestion. The primary reaction sites include the peptide N-terminus and the ε-amino group on lysine side chains. Different samples are encoded with different stable-isotope labels and then mixed for analysis.
Q2: How does Dimethyl labeling calculate relative protein abundance?
During LC-MS/MS analysis, differently isotopically labeled forms of the same peptide have distinguishable mass differences in MS1. Relative peptide ratios are obtained by comparing the peak areas or signal intensities of the corresponding peptide ions. Quantitative information from multiple peptides is then combined to determine relative abundance at the protein level.
Q3: Can Dimethyl labeling provide absolute protein concentrations?
Conventional Dimethyl labeling is primarily used for relative quantification by comparing protein abundance ratios among different samples. If the research objective is to determine the absolute concentration of a specific protein, corresponding standards and a targeted quantitative design are required.
Q4: How many samples can classical Dimethyl labeling compare at one time?
Classical stable-isotope dimethyl labeling can use light, intermediate, and heavy labels for duplex or triplex comparisons. If a project includes additional samples or groups, batch effects associated with a multi-batch design need to be evaluated, or DIA, TMT, and other quantitative strategies should be compared for suitability.
Q5: Why should the sample buffer system be provided in advance?
Dimethyl labeling depends on chemical reactions involving free amino groups. For proteins already extracted or peptides already digested by the customer, buffers, detergents, and other additives may affect labeling or subsequent LC-MS/MS analysis. Complete solution composition is therefore recommended before the project begins.
Q6: What information is recommended before requesting a quotation?
Recommended information includes sample type and species, sample number, group design, biological replicates, available sample amount or protein amount/concentration, buffer system, whether protein extraction or digestion has already been completed, and the primary research objectives. MtoZ Biolabs can use the provided information for free project consultation and study-plan evaluation.
Start Your Project with MtoZ Biolabs
MtoZ Biolabs provides relative quantitative proteomics analysis based on stable-isotope dimethyl labeling and LC-MS/MS. During project consultation, researchers can provide the sample type and species, sample number, group design, biological replicates, available sample amount or protein amount/concentration, buffer system, and research objectives. The technical team can use the provided information to evaluate the Dimethyl labeling strategy, sample preparation, and data-analysis arrangements and provide a project consultation.
MtoZ Biolabs, an integrated chromatography and mass spectrometry (MS) services provider.
Related Services
SILAC/Dimethyl Quantitative Proteomics Service
Label-Based Protein Quantitative Service—iTRAQ, TMT, SILAC
TMT Quantitative Proteomics Analysis Service
