



Liquid chromatography-mass spectrometry (LC-MS), usually as LC-MS/MS, supports quantitative proteomics applications that need relative discovery, closed-list confirmation, or calibrated amount from peptide digests. Common applications include tissue or cell mechanism screens by label-free quantification (LFQ) or data-independent acquisition (DIA), multi-arm panels by tandem mass tags (TMT) or iTRAQ, cultured-cell dynamics by SILAC, biomarker candidate discovery in biofluids, and targeted PRM or MRM confirmation after a shortlist is closed. Absolute applications use matched standards such as AQUA peptides when concentration or stoichiometry is required (Cox et al. 2014; Thompson et al. 2003; Gerber et al. 2003). These uses stay claim-dependent: relative tables are not concentration by default, and discovery lists are not diagnostic cutoffs.
Researchers who already have defined samples, comparison groups, or a specific project objective can also review the MtoZ Biolabs Label-Free Quantitative Proteomics Service, TMT Quantitative Proteomics Analysis Service, and Targeted Proteomics Service for project-specific feasibility and analysis planning.
Application Map Across Quantitative Routes
Application pattern |
LC-MS route often used |
Default claim |
|---|---|---|
Open mechanism or treatment screens |
LFQ with DDA or unlabeled DIA |
Relative same-protein change across independent runs |
Multi-condition panels that fit one kit |
TMT or iTRAQ reporter-ion LC-MS/MS |
Relative ratios inside one plex |
Cultured-cell signaling or drug dynamics |
SILAC metabolic labeling |
Relative ratios in compatible culture systems |
Biofluid biomarker discovery |
LFQ or DIA discovery cohorts |
Candidate lists for later confirmation |
Closed candidate confirmation |
PRM or MRM |
Relative confirmation of scheduled peptides |
Named amount or specification |
AQUA / SIS with PRM or MRM |
Calibrated peptide amount under stated spike timing |
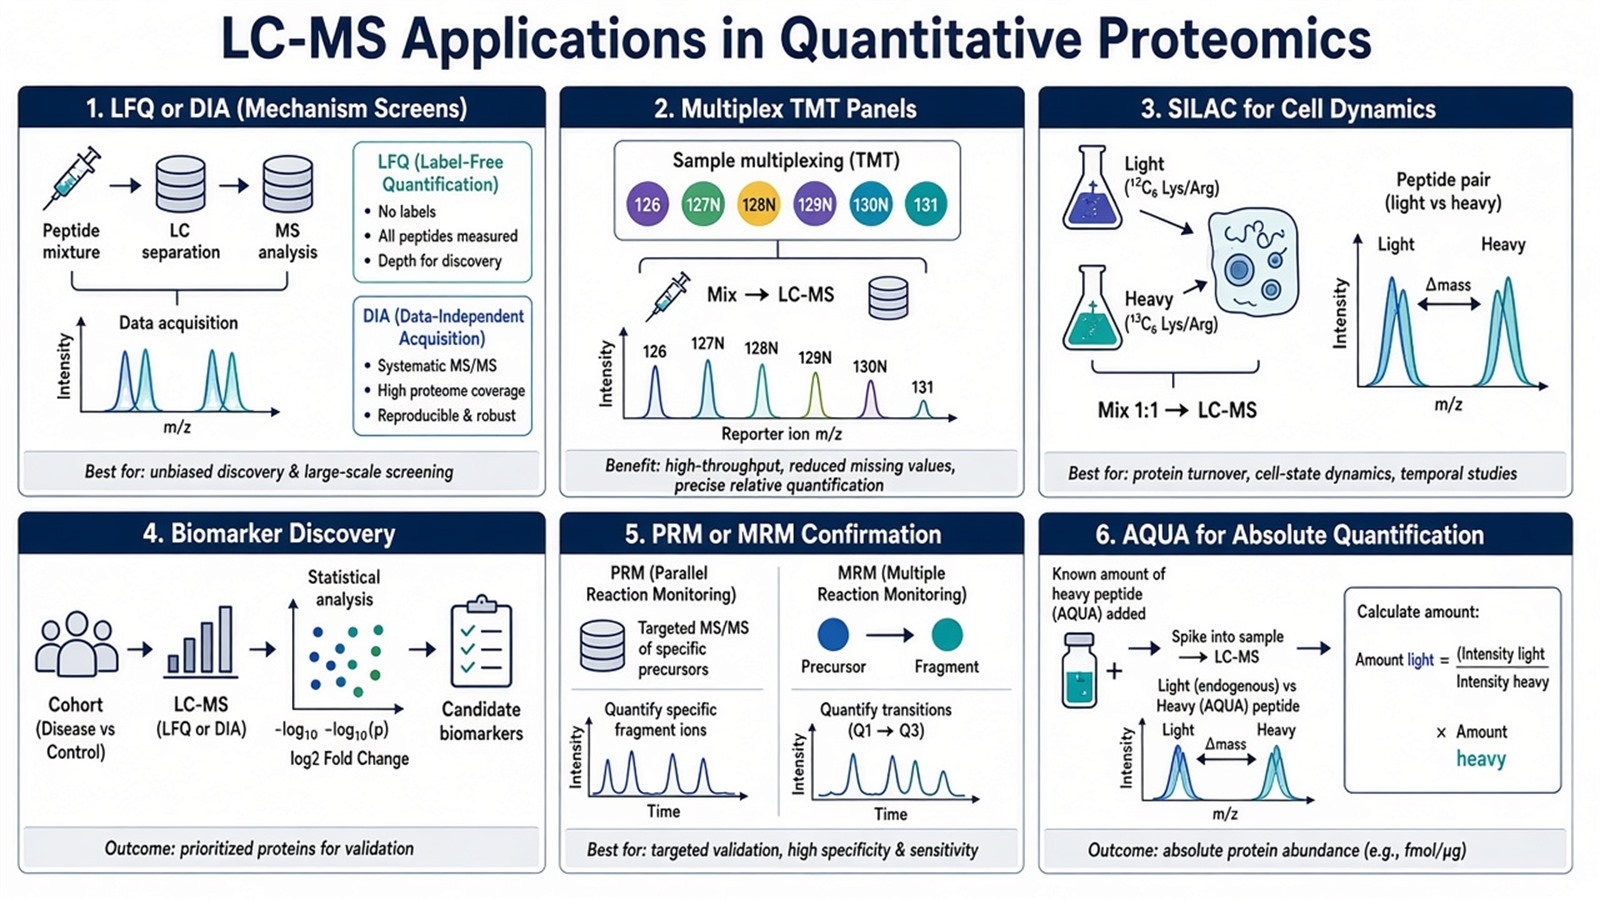
Figure 1. LC-MS applications in quantitative proteomics split by claim: open relative discovery, multiplex or metabolic labeling, closed confirmation, or calibrated absolute amount.
Name the application by the claim it can defend, not by instrument sensitivity alone.
How LC-MS Supports Those Applications
Across routes, liquid chromatography reduces peptide complexity before ionization. Tandem MS identifies peptides from fragment spectra and records quantitative features: MS1 peak areas or DIA fragment chromatograms in LFQ, reporter ions in TMT or iTRAQ, light-to-heavy pairs in SILAC or AQUA, and scheduled transitions or full MS2 in MRM or PRM. The platform is shared; the application outcome is not interchangeable without stating whether samples were labeled, pooled, scheduled, or calibrated.
For LFQ-only application depth, see Applications of LC-MS/MS in Label-Free Quantitative Proteomics. For claim definitions, see Protein Quantification Measured by Mass Spectrometry. For labeled versus unlabeled families, see A Comprehensive Overview of Labeled and Label-Free Quantitative Proteomics Methods.
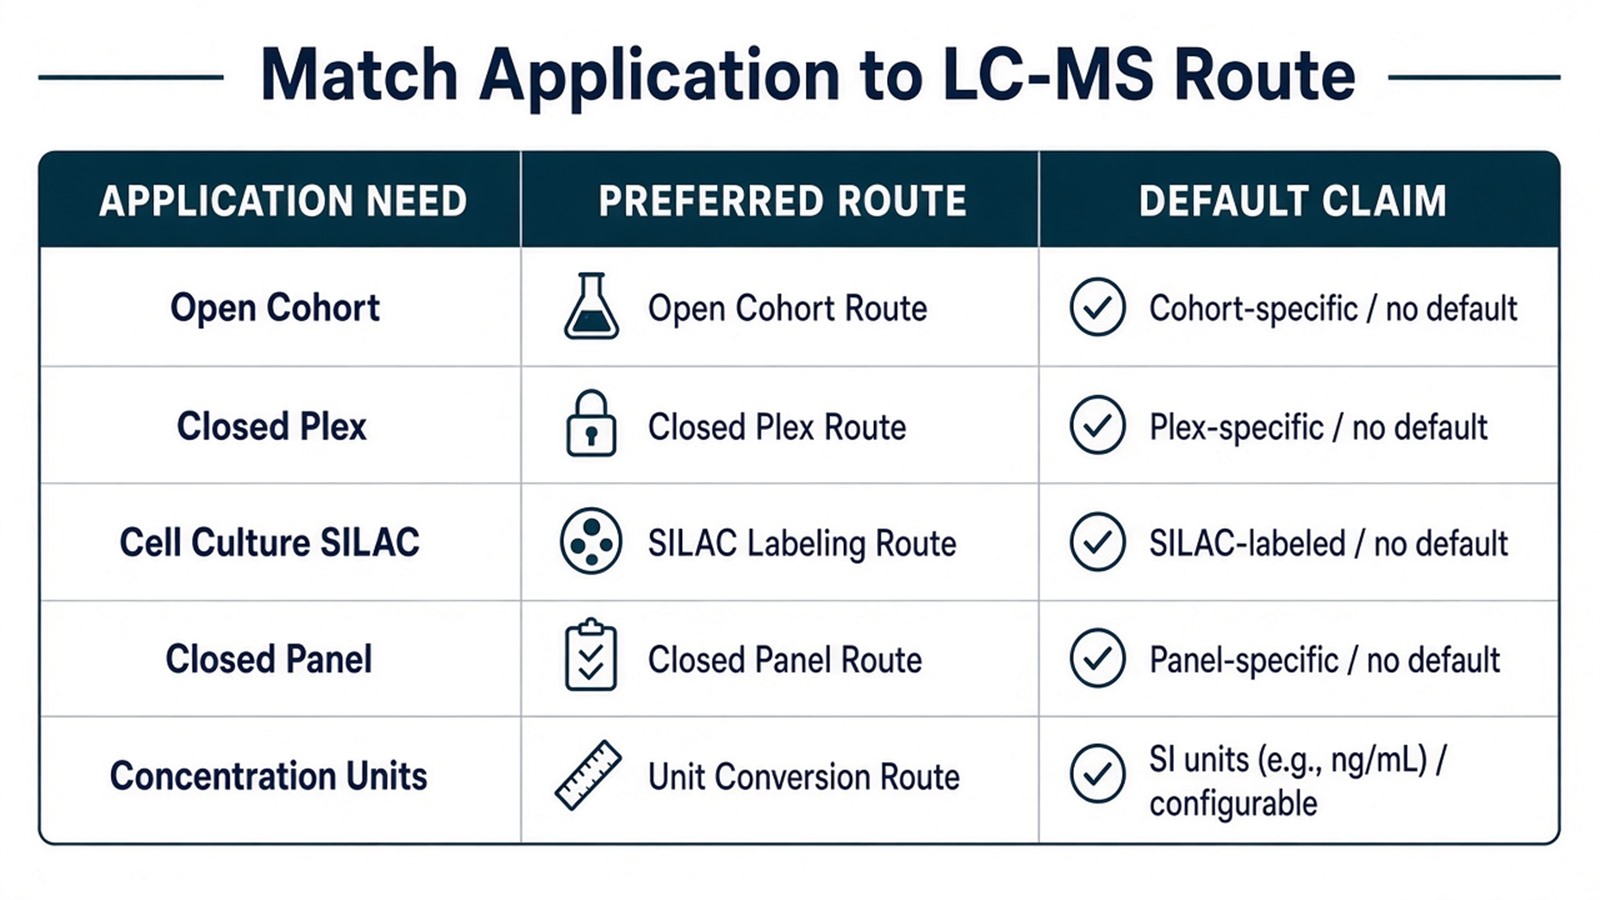
Figure 2. Match the research application to the LC-MS route whose default claim, enrollment rules, and chemistry you can control.
Judgment Criteria: Matching Application to Route
Study need |
Often prefer |
Reassess |
|---|---|---|
Open relative discovery, flexible enrollment |
LFQ or unlabeled DIA |
Absolute units required |
Closed multiplex same-run comparison |
TMT or iTRAQ |
Samples still arriving after mixing |
Metabolic early mix in culture |
SILAC |
Primary tissue or clinical biofluid without culture labeling |
Confirmation of a closed shortlist |
PRM or MRM |
Inventory still open |
Concentration or stoichiometry |
AQUA / SIS absolute assay |
Fold change alone is enough |
Multi-omics integration |
Relative discovery first, then targeted follow-up |
Treating pathway enrichment as clinical proof |
Prefer LFQ or DIA when samples must stay unlabeled. Prefer TMT when every planned arm fits one plex. Prefer SILAC when cultured cells can be labeled metabolically. Prefer PRM or MRM when peptides are closed. Prefer AQUA when the decision sentence needs amount.
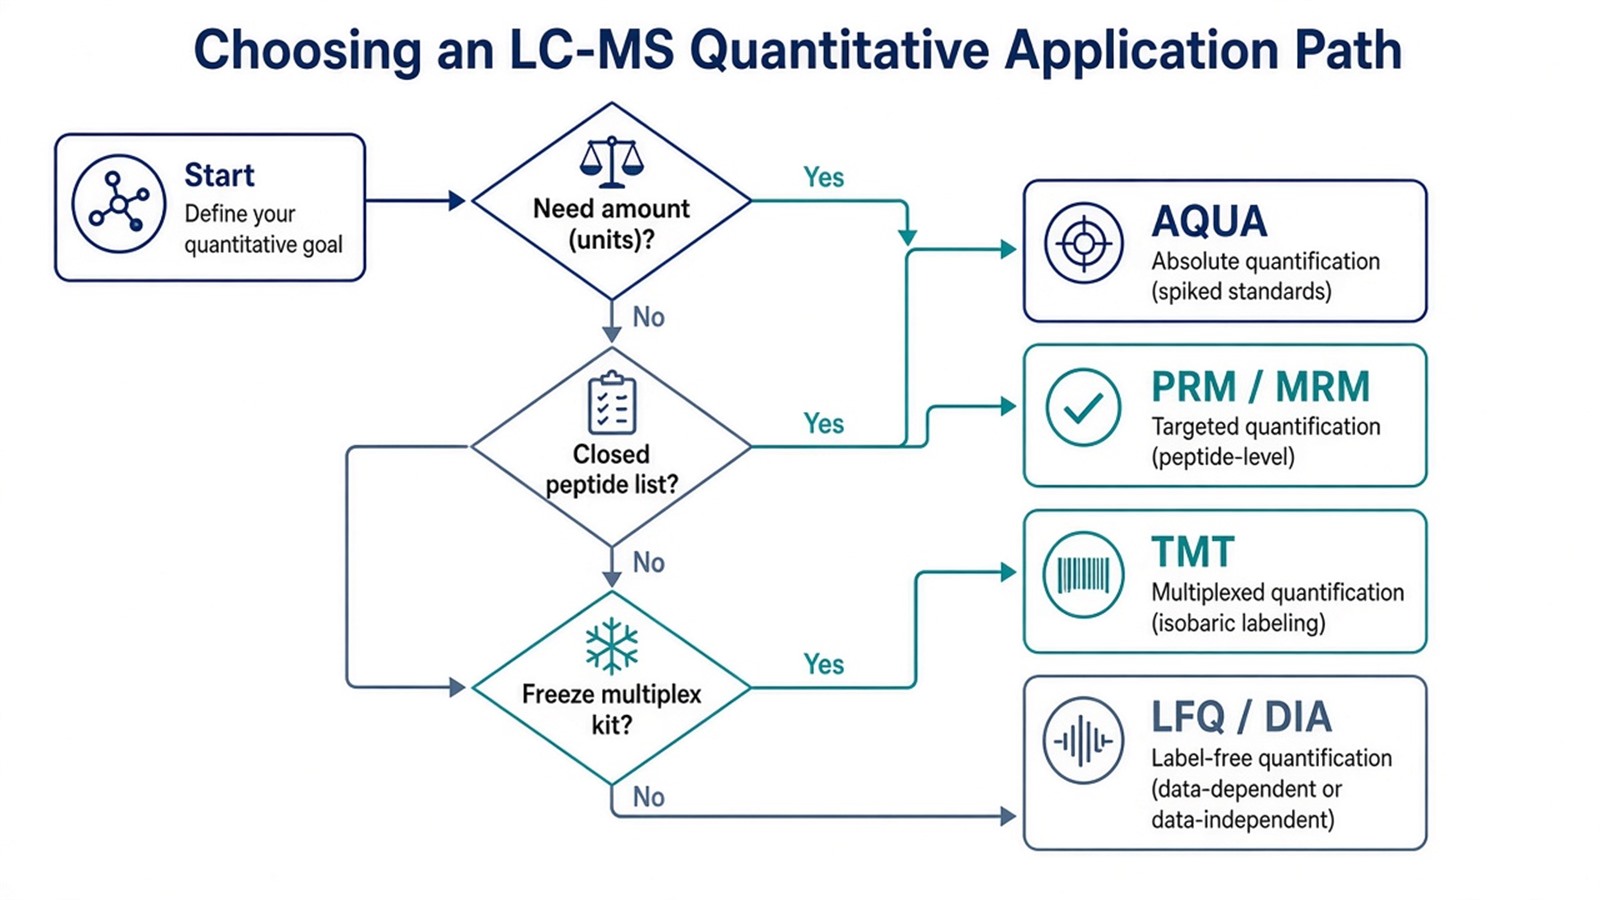
Figure 3. Application selection starts from amount need, panel status, and labeling feasibility, then chooses an LC-MS quantitative path.
For LFQ versus isobaric choice, see Comparing LFQ and Isobaric Labeling Strategies in Proteomics Quantification. For acquisition stacks, see DDA vs. DIA vs. TMT in LC-MS/MS Proteomics: How to Choose the Right Strategy. For DIA limits, see Advantages and Disadvantages of DIA in Protein Quantification. For targeted and absolute trade-offs, see PRM and MRM in Targeted Proteomics: Advantages and Limitations and Advantages and Disadvantages of AQUA Technique.
Limits and Application Boundaries
LC-MS depth does not erase unmatched digestion, confounded run order, ratio compression in isobaric plexes, or chimeric DIA windows. Multi-omics integration still inherits each omics claim; a relative protein list does not prove pathway activity. Single-cell or ultra-low-input LC-MS applications remain constrained by input chemistry and missingness, not by marketing names.
Typical applications with clear boundaries:
- Disease-mechanism and pharmacology screens that need relative proteome change.
- Clinical or preclinical biofluid discovery that later enters PRM or MRM.
- Multi-arm drug or genotype panels inside one TMT kit.
- Cultured-cell SILAC designs for early-mix relative dynamics.
- Named-protein amount when process or research validation needs units.
Diagnostic cutoffs, companion-diagnostic claims, and bulk nutritional protein certificates sit outside ordinary research LC-MS quantitative proteomics reporting unless a separately validated assay framework is in place.
Once the application claim, labeling or unlabeled plan, and panel status are defined, the project can be scoped as LFQ, TMT, targeted, or AQUA work. Researchers can review the MtoZ Biolabs Deep DIA Proteomics Service when unlabeled DIA is part of the application, and the Absolute Quantitative Analysis (AQUA) Service when calibrated amount is required. For sample and pretreatment constraints, see Absolute and Relative Quantitative Proteomics Analysis.
Frequently Asked Questions
1. Is LC-MS the same as LC-MS/MS in these applications?
In quantitative proteomics, applications almost always use tandem MS after LC. LC-MS alone rarely delivers peptide identification and quantification together for discovery cohorts.
2. Can one LC-MS method cover every quantitative application?
No. Open discovery, multiplex labeling, closed confirmation, and absolute amount use different designs and QC.
3. Does DIA replace TMT for clinical cohorts?
Not automatically. DIA keeps samples unlabeled and independent. TMT freezes a multiplex kit for same-run ratios. Choose from enrollment and chemistry, not from a universal ranking.
4. When should an application move from discovery LC-MS to targeted LC-MS?
When the protein inventory is closed enough to schedule proteotypic peptides for confirmation or amount.
5. Do LC-MS quantitative applications provide clinical cutoffs by default?
No. Ordinary research tables need a separately validated assay framework before diagnostic decisions.
Reference
- J. Cox, M.Y. Hein, C.A. Luber, et al. (2014). Accurate proteome-wide label-free quantification by delayed normalization and maximal peptide ratio extraction, termed MaxLFQ. Mol. Cell. Proteomics, 13, 2513-2526. https://doi.org/10.1074/mcp.M113.031591
- A. Thompson, J. Schaefer, K. Kuhn, S. Kienle, J. Schwarz, G. Schmidt, T. Neumann, C. Hamon (2003). Tandem mass tags: a novel quantification strategy for comparative analysis of complex protein mixtures by MS/MS. Anal. Chem., 75, 1895-1904. https://doi.org/10.1021/ac0262560
- S.A. Gerber, J. Rush, O. Stemman, M.W. Kirschner, S.P. Gygi (2003). Absolute quantification of proteins and phosphoproteins from cell lysates by tandem MS. Proc. Natl. Acad. Sci. U.S.A., 100, 6940-6945. https://doi.org/10.1073/pnas.0832254100
