



Quantitative proteomics compares protein abundance with mass spectrometry (MS). Label-free quantification (LFQ) keeps unlabeled samples as independent liquid chromatography-tandem mass spectrometry (LC-MS/MS) runs and rebuilds relative values by alignment and normalization. Tandem mass tag (TMT) and isobaric tags for relative and absolute quantitation (iTRAQ) are isobaric labeling routes: peptides from each channel are tagged, pooled, and quantified from reporter-ion ratios after fragmentation (Thompson et al. 2003; Ross et al. 2004; Cox et al. 2014). All three are widely used relative strategies, not calibrated absolute concentration by default. LFQ fits open cohorts and flexible enrollment. TMT and iTRAQ fit closed multiplex designs that need same-run comparison when chemistry and channel count match the study plan.
Researchers who already have defined samples, comparison groups, or a specific project objective can also review the MtoZ Biolabs Label-Free Quantitative Proteomics Service and TMT Quantitative Proteomics Analysis Service for project-specific feasibility and analysis planning.
Label-Free, TMT, and iTRAQ at a Glance
Route |
Labeling |
Sample handling |
Quantitative readout |
Typical fit |
|---|---|---|---|---|
Label-free (LFQ) |
None |
Independent injections |
MS1 MaxLFQ or unlabeled DIA fragments |
Large or evolving cohorts, labeling avoided |
TMT |
Isobaric amine tags |
Pooled multiplex plex |
Reporter ions at MS2 or MS3 |
Multi-arm panels that fit one TMT kit |
iTRAQ |
Isobaric amine tags (same class as TMT) |
Pooled multiplex plex |
Reporter ions after fragmentation |
Same strategic role as TMT with kit-specific plex size |
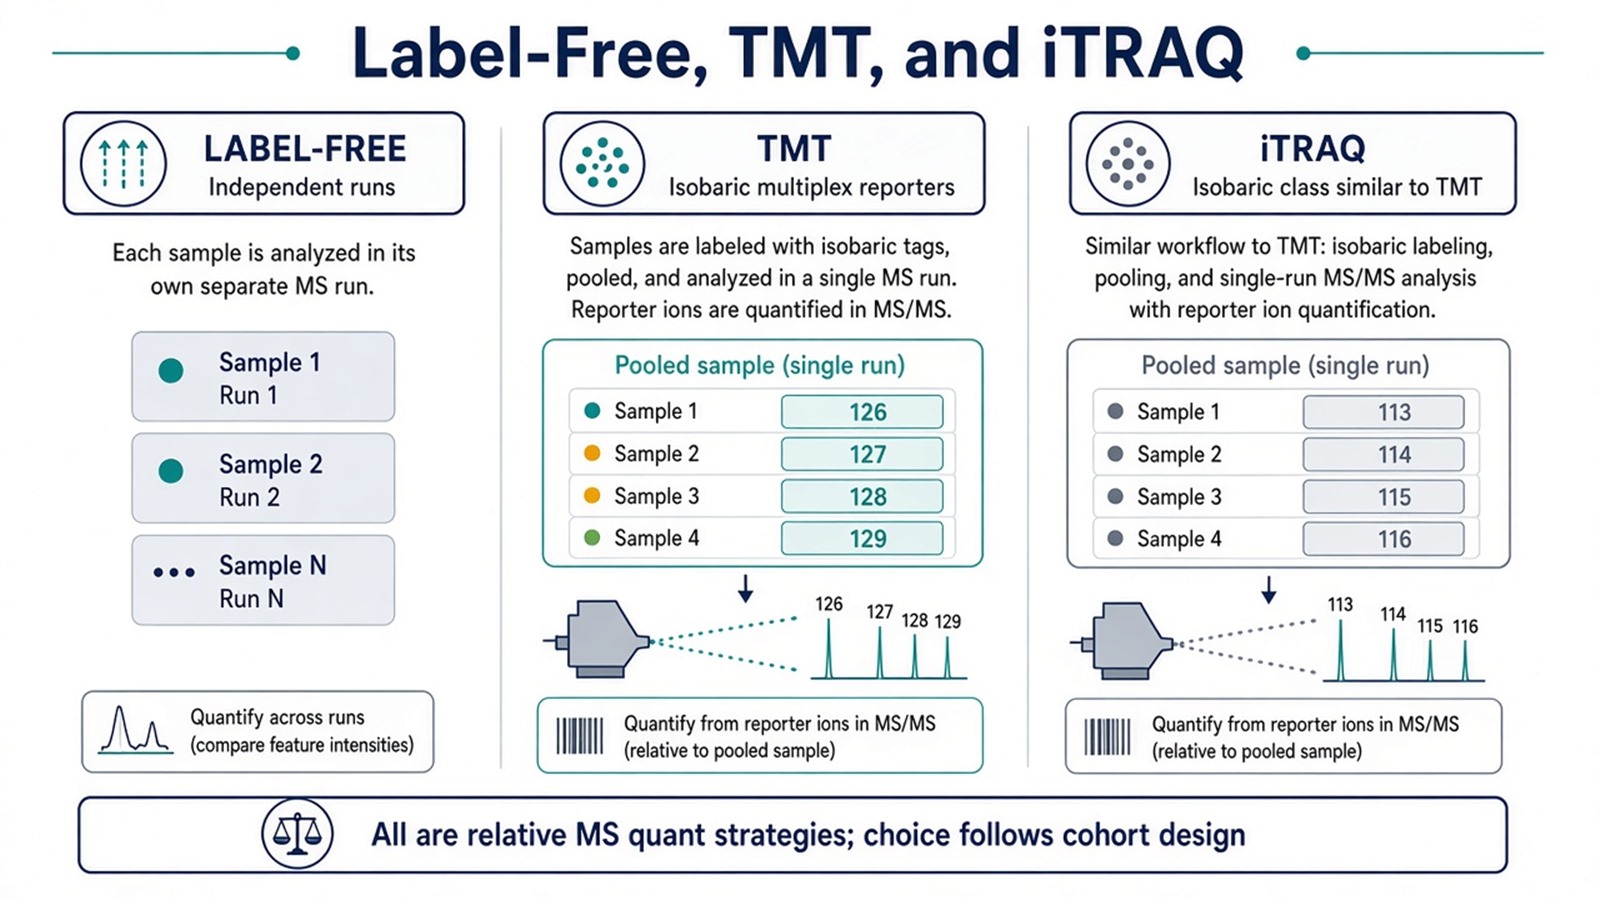
Figure 1. LFQ, TMT, and iTRAQ are all MS-based relative quant routes; they differ in labeling, mixing rules, and cohort closure.
Pick the route whose main constraint you can control: unlabeled flexibility, or a closed isobaric plex with reporter readout.
TMT Quantitative Proteomics: Principle and Workflow
TMT reagents carry an amine-reactive group, a mass balancer, and a channel-specific reporter. After tryptic digestion, each sample receives one tag, channels are combined, and the mixed sample is analyzed by LC-MS/MS. At MS1, labeled forms of the same peptide appear as one isobaric precursor. Upon data-dependent acquisition (DDA) fragmentation, reporter ions of distinct masses report relative channel intensities. Software integrates reporters into peptide and protein ratios inside that plex.
A typical TMT workflow runs in order: extract and digest proteins under one protocol, label each channel and quench, remove free tag, pool labeled peptides, optionally fractionate to reduce complexity, acquire LC-MS/MS with reporter readout at MS2 or with synchronous precursor selection MS3 (SPS-MS3) on compatible platforms, then search spectra and summarize ratios with false discovery rate (FDR) control.
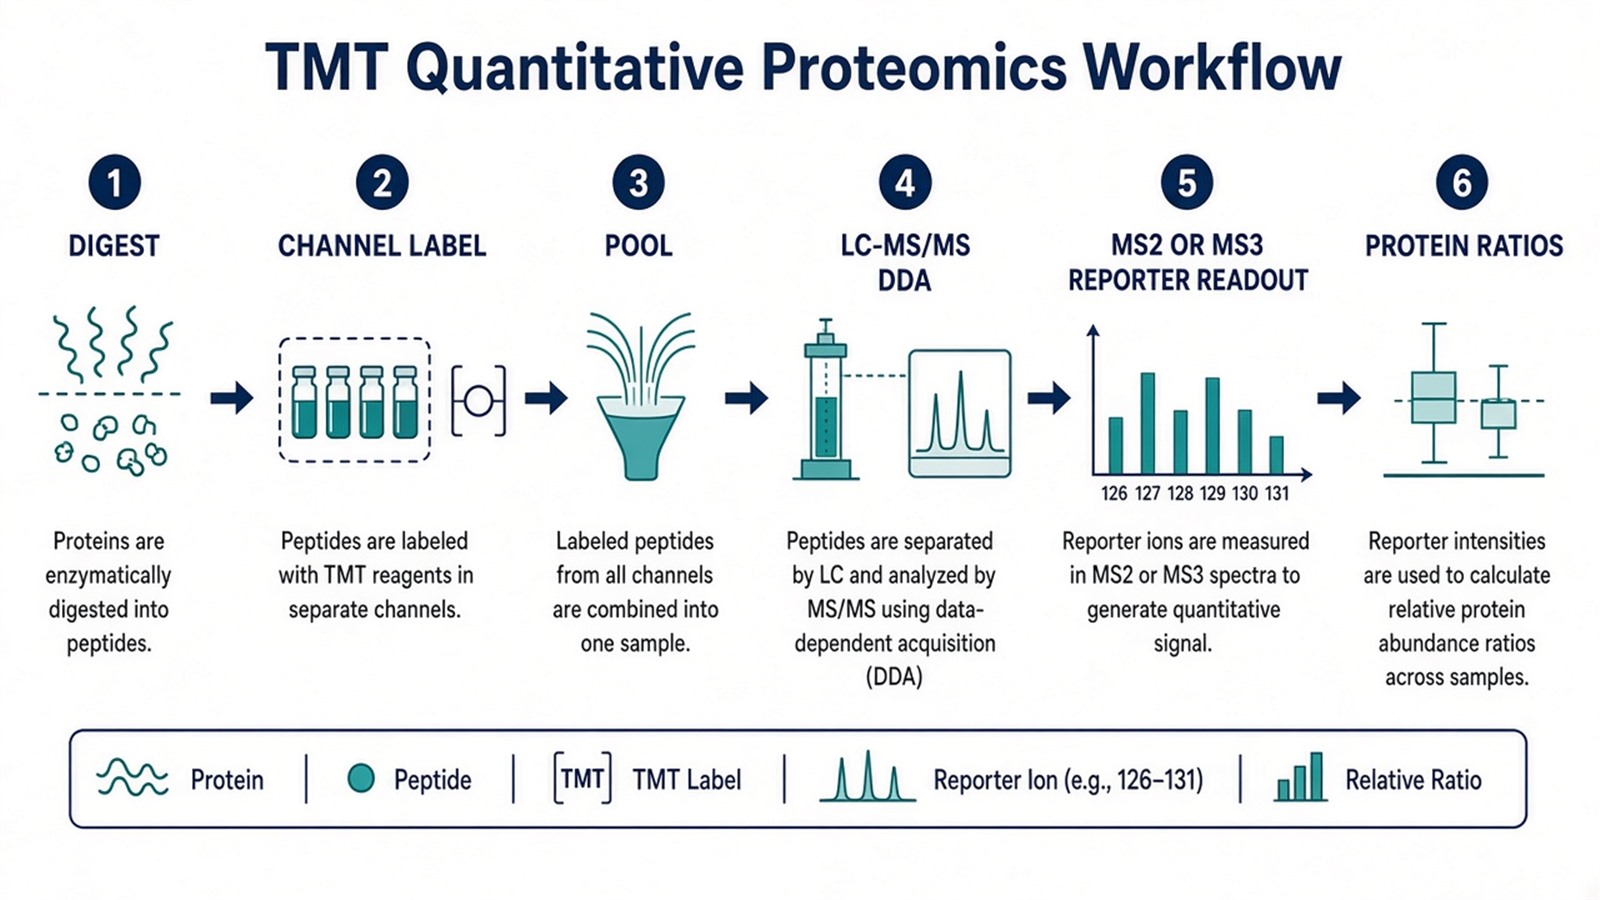
Figure 2. TMT quantification is defined by reporter-ion readout after labeling and pooling, not by MS1 precursor areas across independent runs.
For reporter-ion definitions, see Introduction to TMT Reporter Ions and Mass Spectrometry Analysis. For labeling chemistry detail, see Mechanism of TMT Labeling in Mass Spectrometry. For the full experimental sequence, see A Detailed Workflow of TMT-Based Quantitative Proteomics.
TMT Judgment Criteria, Limits, and Applications
Study need |
TMT often fits |
Reassess before labeling |
|---|---|---|
Design |
Closed channel map before mixing |
Samples still arriving after pool |
Chemistry |
Clean amine labeling at N-termini and lysines |
Blocking buffers or failed digestion |
Claim |
Same-run relative ratios inside one plex |
Absolute concentration or diagnostic cutoff |
Acquisition |
DDA MS2 or planned MS3 reporter integration |
Treating unlabeled DIA as TMT readout |
Scale |
Groups and replicates fit one commercial kit |
Sample count forces many plexes without bridging |
TMT limits include finite channel count, ratio compression when co-isolated peptides mix reporters at MS2, incomplete labeling, and the need for bridging design across separate plexes. MS3 or SPS-MS3 can reduce compression but does not replace sound plex layout or absolute quantification claims.
Typical TMT applications include multi-condition drug or genotype panels, dose-response arms inside one kit, and biomarker discovery lists that need relative ranking within a frozen plex. Diagnostic cutoffs and regulatory assay claims sit outside ordinary research TMT reporting unless a separately validated framework is in place.
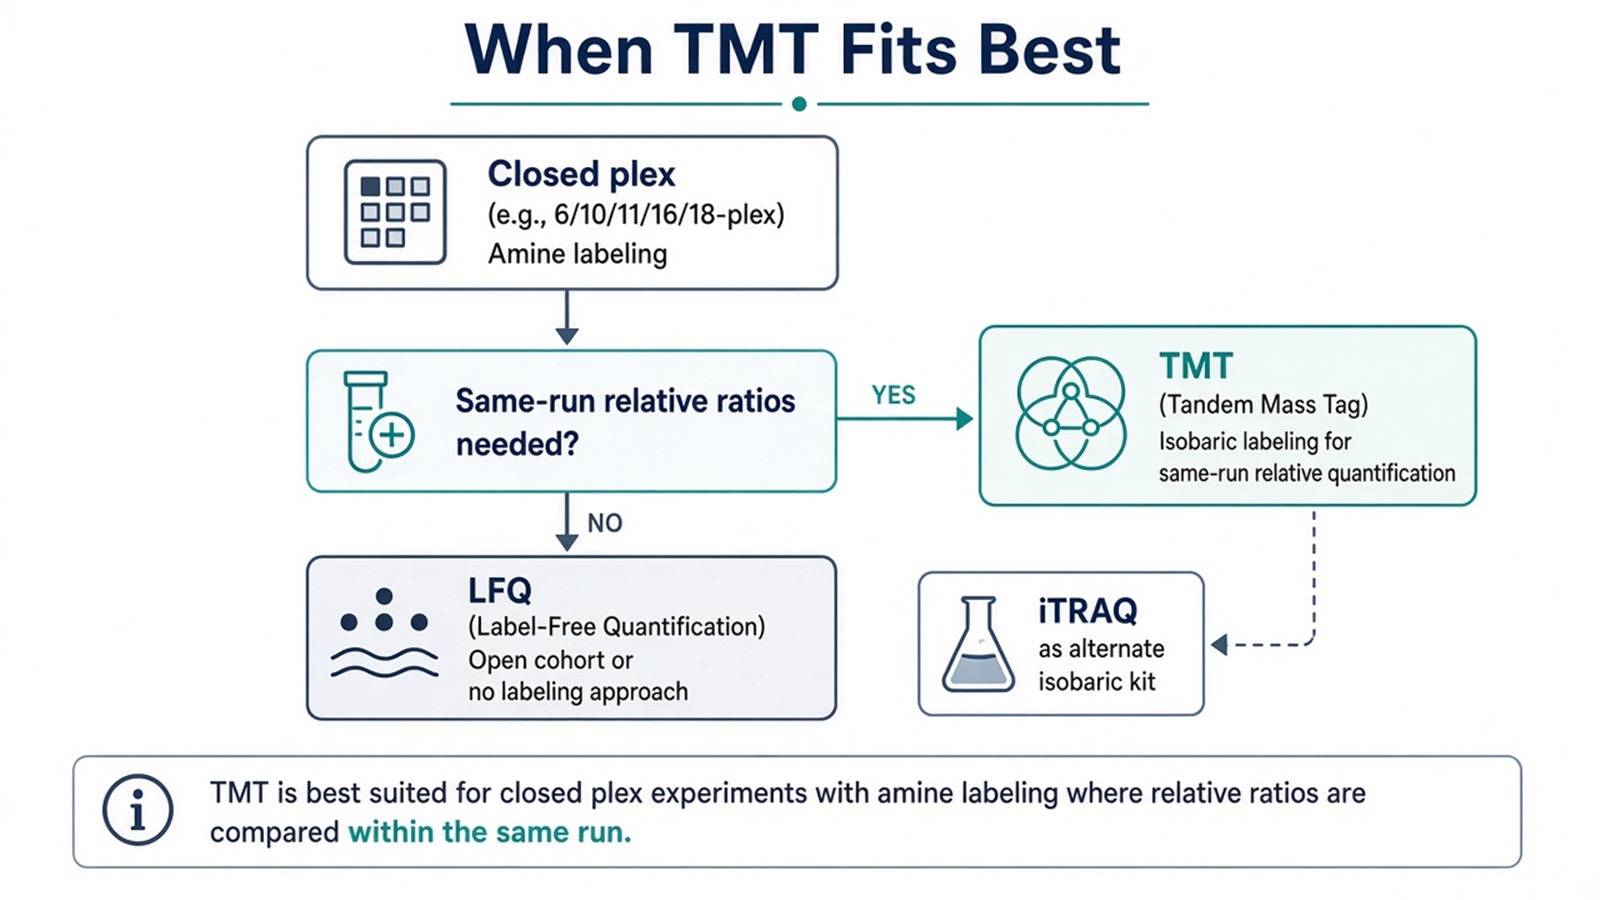
Figure 3. TMT fits closed relative multiplex designs; LFQ fits open unlabeled cohorts; iTRAQ follows the same isobaric rules with kit-specific plex limits.
For compression mechanism, see How Does Ratio Compression Arise in TMT-Based Quantitative Proteomics and Why Is It Limited to Relative Quantification?. For MS3 mitigation, see How Can SPS-MS3 Be Utilized to Optimize TMT Workflows and Minimize Ratio Compression?. For TMT versus unlabeled selection, see TMT vs Label Free Quantitative Proteomics and Comparing LFQ and Isobaric Labeling Strategies in Proteomics Quantification. For acquisition stacks, see DDA vs. DIA vs. TMT in LC-MS/MS Proteomics: How to Choose the Right Strategy.
Where Label-Free and iTRAQ Sit in the Same Picture
LFQ omits tags and accepts run-to-run drift control through randomization, QC pools, and normalization. It pairs with DDA-LFQ or unlabeled DIA inside the same unlabeled strategy. iTRAQ belongs to the isobaric class alongside TMT: pooled channels, reporter readout, closed enrollment, and similar co-isolation limits. Kit plex size and vendor chemistry differ, but the strategic choice between LFQ and isobaric labeling follows the same cohort rules.
For LFQ workflow detail, see What Is Label-Free Analysis? LFQ Principles, Workflow, Advantages, and Limitations. For design trade-offs, see Advantages and Disadvantages of Label-Free Quantitative Proteomics and Advantages and Disadvantages of TMT-Based Quantitative Proteomics Analysis.
Once the route, channel map if isobaric, and relative claim are defined, the project can be scoped as LFQ, TMT, or iTRAQ-class labeling. Researchers can review the MtoZ Biolabs TMT Quantitative Proteomics Analysis Service and Label-Free Quantitative Proteomics Service.
Frequently Asked Questions
1. Are TMT and iTRAQ interchangeable with label-free quantification?
No. LFQ uses independent unlabeled runs. TMT and iTRAQ pool labeled channels and read reporter ratios in one plex.
2. Does TMT provide absolute protein quantification?
Not by default. TMT reporters support relative comparison inside one plex unless matched standards and calibration are added separately.
3. Is TMT always more accurate than LFQ?
Neither is universally better. TMT can reduce injection drift among channels in the same run. LFQ can scale sample count and open enrollment when QC is controlled.
4. What is the main TMT quantitative risk?
Co-isolation at MS2 can compress fold changes. Fractionation, MS3, and plex design manage that risk; they do not remove it entirely.
5. When should I choose iTRAQ instead of TMT?
When the available isobaric kit and plex size match the cohort and the same closed multiplex rules apply. Strategic choice still starts from labeling versus label-free, not brand alone.
Reference
- A. Thompson, J. Schaefer, K. Kuhn, S. Kienle, J. Schwarz, G. Schmidt, T. Neumann, C. Hamon (2003). Tandem mass tags: a novel quantification strategy for comparative analysis of complex protein mixtures by MS/MS. Anal. Chem., 75, 1895-1904. https://doi.org/10.1021/ac0262560
- P.L. Ross, Y.N. Huang, J.N. Marchese, et al. (2004). Multiplexed protein quantitation in Saccharomyces cerevisiae using amine-reactive isobaric tagging reagents. Mol. Cell. Proteomics, 3, 1154-1169. https://doi.org/10.1074/mcp.M400049-MCP200
- J. Cox, M.Y. Hein, C.A. Luber, et al. (2014). Accurate proteome-wide label-free quantification by delayed normalization and maximal peptide ratio extraction, termed MaxLFQ. Mol. Cell. Proteomics, 13, 2513-2526. https://doi.org/10.1074/mcp.M113.031591
