



Tandem mass tag (TMT) quantitative proteomics compares relative protein abundance after peptides from different samples are chemically labeled, mixed, and acquired in one liquid chromatography-tandem mass spectrometry (LC-MS/MS) run. Reporter ions released during fragmentation carry the quantitative comparison. The practical advantage is that injection-to-injection chromatography drops out of that comparison for samples that share one multiplex set. The practical cost is that fold changes read from mixed reporter ions can be compressed, the cohort must fit a labeling kit, and ordinary TMT intensities are not protein amounts. Advantages listed as high throughput or high sensitivity only hold after labeling chemistry, plex structure, and the relative-comparison claim are specified.
Researchers who already have defined samples, comparison groups, or a specific project objective can also review the MtoZ Biolabs TMT Quantitative Proteomics Analysis Service for project-specific feasibility and analysis planning.
Why TMT Trade-offs Appear at Experimental Design
A typical project reaches TMT when several conditions, time points, or biological replicates need to be compared together, and when the laboratory is willing to freeze that set before labeling. The scientific question has shifted from whether proteins can be identified to whether reporter-ion ratios inside one pooled injection can support the intended contrast.
TMT is an isobaric labeling method (Thompson et al. 2003). Acquisition is a separate layer. In MtoZ isobaric-labeling offerings, TMT workflows are typically acquired in data-dependent acquisition (DDA) so that reporter ions can be read. Data-independent acquisition (DIA) is not a TMT synonym, and unlabeled discovery remains a different sample-handling choice.
If samples cannot enter a complete labeling reaction, arrive after the plex is closed, or require an amount rather than a ratio, the listed TMT advantages do not transfer.
How TMT Reporter Ions Create a Relative Comparison
Proteins are extracted and digested, usually with trypsin. Each digest is labeled on peptide N-termini and lysine residues with a TMT reagent from one multiplex kit. Labeled peptides are combined into a single pool, separated by liquid chromatography, and fragmented. At the MS1 level the tags are isobaric, so the same peptide from different channels co-elutes as one precursor. Upon fragmentation, reporter ions of distinct masses appear, and their intensities are used to infer relative peptide, then protein, abundance across channels.
MS2 reporter ions are recorded from the isolated precursor window. Co-isolated peptides can contribute reporters and pull ratios toward one, an effect known as ratio compression (Ting et al. 2011). Some tribrid methods, including synchronous precursor selection MS3 (SPS-MS3), select fragment ions before a second fragmentation so that reporters are read from a cleaner ion population (McAlister et al. 2014). MS3 reduces compression. It does not create an absolute concentration, and it does not recover a sample that was labeled incompletely.
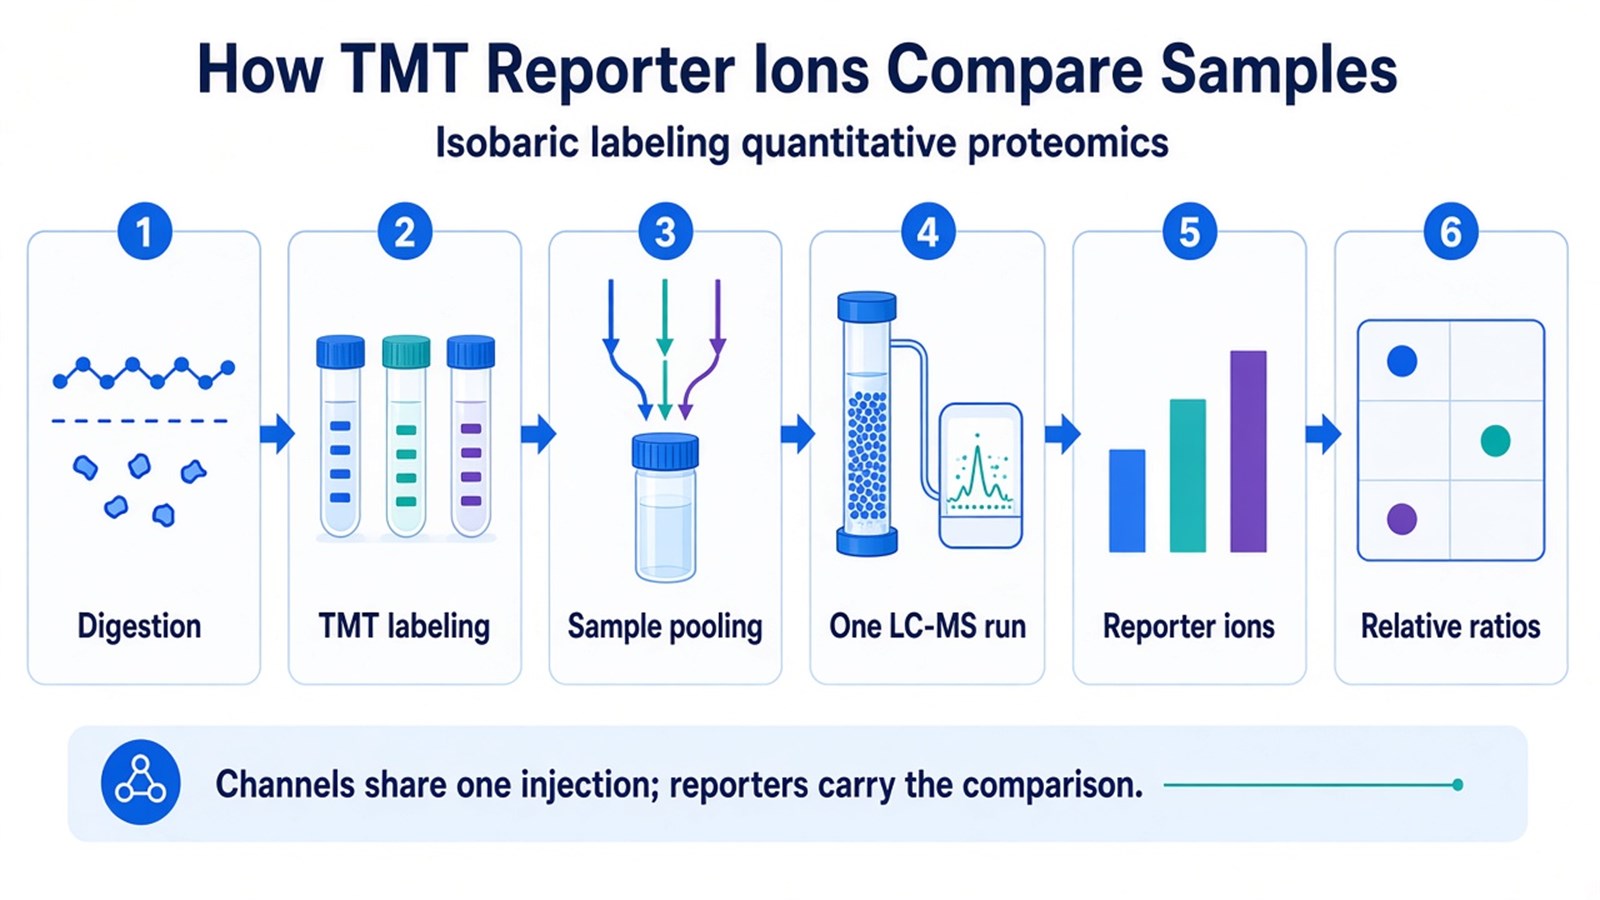
Figure 1. TMT compares channels inside one pooled LC-MS/MS run; reporter ions, not MS1 peak areas of separate injections, carry the quantitative ratio.
A TMT value is a relative reporter-ion ratio inside one multiplex set, not a calibrated protein amount.
Advantages That Hold After Samples Can Be Labeled and Pooled
The usable advantages of TMT are conditional. They appear when every sample in the contrast can be labeled with the same kit, mixed, and acquired under one chromatographic sequence.
Same-run comparison across a multiplex set
Because labeled peptides share one injection, differences in column aging, spray stability, and run order between separately injected unlabeled files are no longer the main comparison noise. That is the design reason TMT is used for multi-group relative discovery. It is not proof that labeling chemistry is unbiased, or that incomplete labeling will be invisible in the ratios.
Commercially available TMT kits support multiplex sets with different channel counts, including formats used as 6-plex through 18-plex depending on the reagent. The useful set is the one that holds the planned groups and replicates without forcing unrelated samples into leftover channels.
Time-course and multi-condition designs that fit one kit
Dose series, treatment-versus-control panels, and small factorial designs can sit inside one plex when group definitions are closed before labeling. Biological replicates can occupy parallel channels, which keeps the statistical contrast inside the same mixed run. A design that still expects later enrollment, extra tissues, or an open-ended cohort does not keep that advantage, because a new kit becomes a new analytical block.
Fewer missing identifications than independently injected DDA-LFQ
In unlabeled data-dependent acquisition, a peptide may be identified in one file and missed in another. After TMT mixing, identification evidence is collected from the pooled precursor, so the quantitative matrix is often more complete across channels in that run. Completeness is not the same as accuracy. A peptide present in every channel can still carry a compressed ratio if co-isolation is severe.
Unlabeled label-free quantification remains the better match when samples must stay as independent injections.
When sample number, multiplexing, or later enrollment makes that choice less straightforward, researchers can refer to Comparing LFQ and Isobaric Labeling Strategies in Proteomics Quantification for a more detailed strategy-selection framework.
Limitations That Change the Experimental Plan
TMT disadvantages are design constraints. They decide whether remaining material should enter a labeling reaction at all.
Ratio compression at MS2
Co-isolated precursors contribute reporter ions in the same MS2 scan, which underestimates true fold changes, especially in complex tissue or biofluid digests (Ting et al. 2011). Prefractionation can lower co-isolation by reducing peptide density in each run. SPS-MS3 can further reduce reporter mixing on instruments that support that scan type (McAlister et al. 2014). Neither step guarantees that a small biological change will remain distinguishable, and both add acquisition or fractionation load.
Relative ratios, not amounts
Reporter intensities support same-peptide comparison across channels in the plex. They do not report copies per cell or a tissue concentration. Ionization efficiency still differs among peptides, so protein A cannot be ranked against protein B by reporter height. An amount claim needs a matched standard and a calibration model, which is a targeted experiment rather than a standard TMT discovery table.
Labeling chemistry, plex size, and later enrollment
TMT reagents require peptides that can be labeled at amines and cleaned of excess tag. Residual detergent, amine buffers, or incomplete digestion can leave unlabeled or partially labeled peptides that distort ratios. Channel count is finite. Samples that arrive after the set is mixed cannot be inserted by ordinary normalization. A second TMT set is a new batch, not an extension of the first ratio matrix.
Reagent cost scales with plex size and with how many extra channels are purchased to hold unused groups. That cost is a planning input. It is not evidence that unlabeled discovery is scientifically weaker.
Post-translational modification studies are not blocked by TMT labeling itself, because labeling occurs after digestion. Site-level quantification still needs enrichment of the modified peptides. Global protein ratios and site occupancy should not be read as the same number.
How to Judge Whether TMT Fits the Current Cohort
The useful test is whether the contrast can be frozen inside one labeling set, whether peptides can be labeled completely, and whether a relative ratio is the claim the paper needs.
Study condition |
TMT is often useful |
Reassess before labeling |
|---|---|---|
Comparison structure |
Groups and replicates fit one multiplex kit |
Enrollment would continue after the plex is mixed |
Quantitative claim |
Same-protein relative change across channels |
Amount or copy number without a matched standard |
Sample chemistry |
Digests can be labeled at amines and cleaned |
Amine buffers or detergents still block labeling |
Missingness vs compression |
A more complete matrix is worth compressed fold changes |
True fold-change magnitude is the main readout |
Adjacent option |
Closed multi-group relative discovery |
Independent injections or a predefined peptide panel |
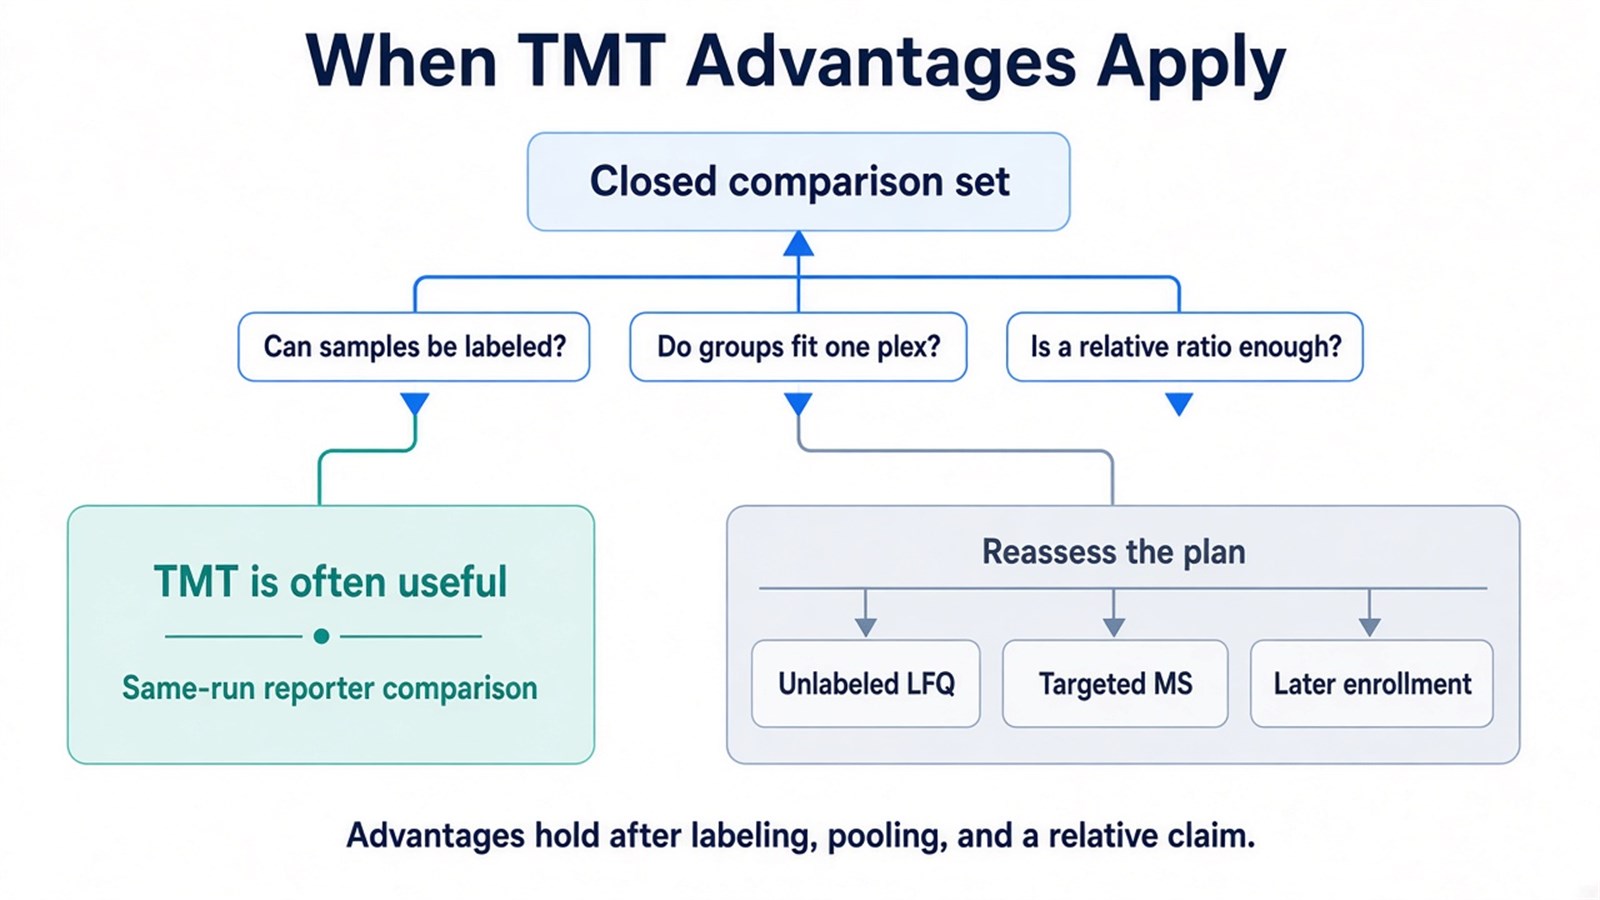
Figure 2. TMT advantages apply when samples can be labeled, pooled, and compared as relative reporter-ion ratios inside one multiplex set.
Frequently Asked Questions
1. Is TMT the same method as iTRAQ?
Both are isobaric peptide tags that release reporter ions after fragmentation. They share the same-run comparison logic and the same co-isolation problem. Kit chemistry, available channel counts, and current laboratory practice differ. Naming one tag does not specify the acquisition mode.
2. Does TMT report absolute protein concentration?
Not from ordinary reporter-ion ratios. Those ratios compare channels inside the plex. Concentration or copy-number claims need a matched standard and a calibration model, usually in a targeted assay.
3. Does MS3 remove ratio compression?
SPS-MS3 can reduce reporter mixing by reading tags after an extra isolation and fragmentation step. Compression can remain in dense regions of a complex digest. Fractionation, isolation width, and sample complexity still belong in the method record.
4. When is unlabeled LFQ the better next experiment than TMT?
Unlabeled LFQ is often the better next experiment when samples cannot be labeled cleanly, when the cohort cannot be frozen inside one kit, or when later samples must be injected independently. TMT is often the better next experiment when the contrast is closed, labeling is feasible, and same-run relative comparison is the main need.
Once the group structure, labeling feasibility, and expected protein-level readout are defined, the project can be evaluated as a TMT study. Researchers can review the MtoZ Biolabs TMT Quantitative Proteomics Analysis Service for project-specific sample evaluation, feasibility assessment, and workflow planning.
