



A rat cell pellet enters protein mutation analysis when the laboratory needs peptide-level evidence that an amino acid substitution is present in the expressed proteome. Bottom-up liquid chromatography-tandem mass spectrometry (LC-MS/MS) digests rat cell proteins, records precursor and fragment spectra, and matches those spectra to a Rattus norvegicus sequence set. A mutation claim at the protein level is supported only when a variant peptide is accepted under declared search settings. A DNA edit, a Western blot band shift, or a pathway annotation does not substitute for that peptide evidence.
Species database choice, expected mutant sequences, digestion coverage, and peptide uniqueness decide which substitutions can be reported. Post-translational modifications change mass for different chemical reasons and must be parameterized separately from sequence variants.
What Protein Mutation Analysis Means in Rat Cell Proteomics
In this setting, protein mutation analysis means detecting an amino acid substitution, insertion, or deletion as a sequence variant in peptides from rat cell extracts. The analytical object is a peptide-spectrum match whose precursor mass and fragment pattern fit the variant sequence better than the wild-type sequence under a false discovery rate (FDR) or a targeted confirmation rule.
Wild-type and variant peptides differ by the mass shift of the substituted residue and by fragment ions that localize the change when coverage allows. Shared peptides that do not contain the mutated site cannot distinguish alleles. Isobaric residues such as leucine and isoleucine keep the same nominal mass in ordinary MS/MS and may remain ambiguous without an orthogonal method.
Gene editing can introduce a coding change in the rat genome. Mass spectrometry asks whether the corresponding protein product yields a detectable variant peptide in the submitted extract. Those are sequential questions, not interchangeable assays.
A protein mutation claim from rat cells requires accepted variant-peptide evidence in a rat-compatible search design.
Researchers who need how sequence and mutation work is structured for protein or peptide samples can refer to the MtoZ Biolabs Sequence and Mutation Analysis Service of Unknown Proteins or Peptides.
How Rat Cell Samples Are Turned into Variant Peptide Evidence
The usual order is fixed: a washed rat cell pellet, lysis, digestion, LC-MS/MS, database search with a rat reference and any declared mutant sequences, then review of wild-type versus variant peptide support. Residual medium, incomplete lysis, and wrong species databases change which peptides appear before any mutation sentence is written.
For known engineered mutants, the expected amino acid change should be added to the search database as a custom FASTA entry or as an explicit variant rule. Searching only a standard UniProt rat proteome will miss substitutions that are absent from that reference. For unknown substitutions on a purified protein or peptide, multi-enzyme mapping and de novo interpretation may be required; a complex whole-cell digest is a harder place to start when the mutation site is undefined.
Targeted parallel reaction monitoring (PRM) or multiple reaction monitoring (MRM) fits when the variant peptide sequence is already closed and needs scheduled confirmation. Open discovery fits when the variant list is still being built and a searchable sequence set can include the candidates.
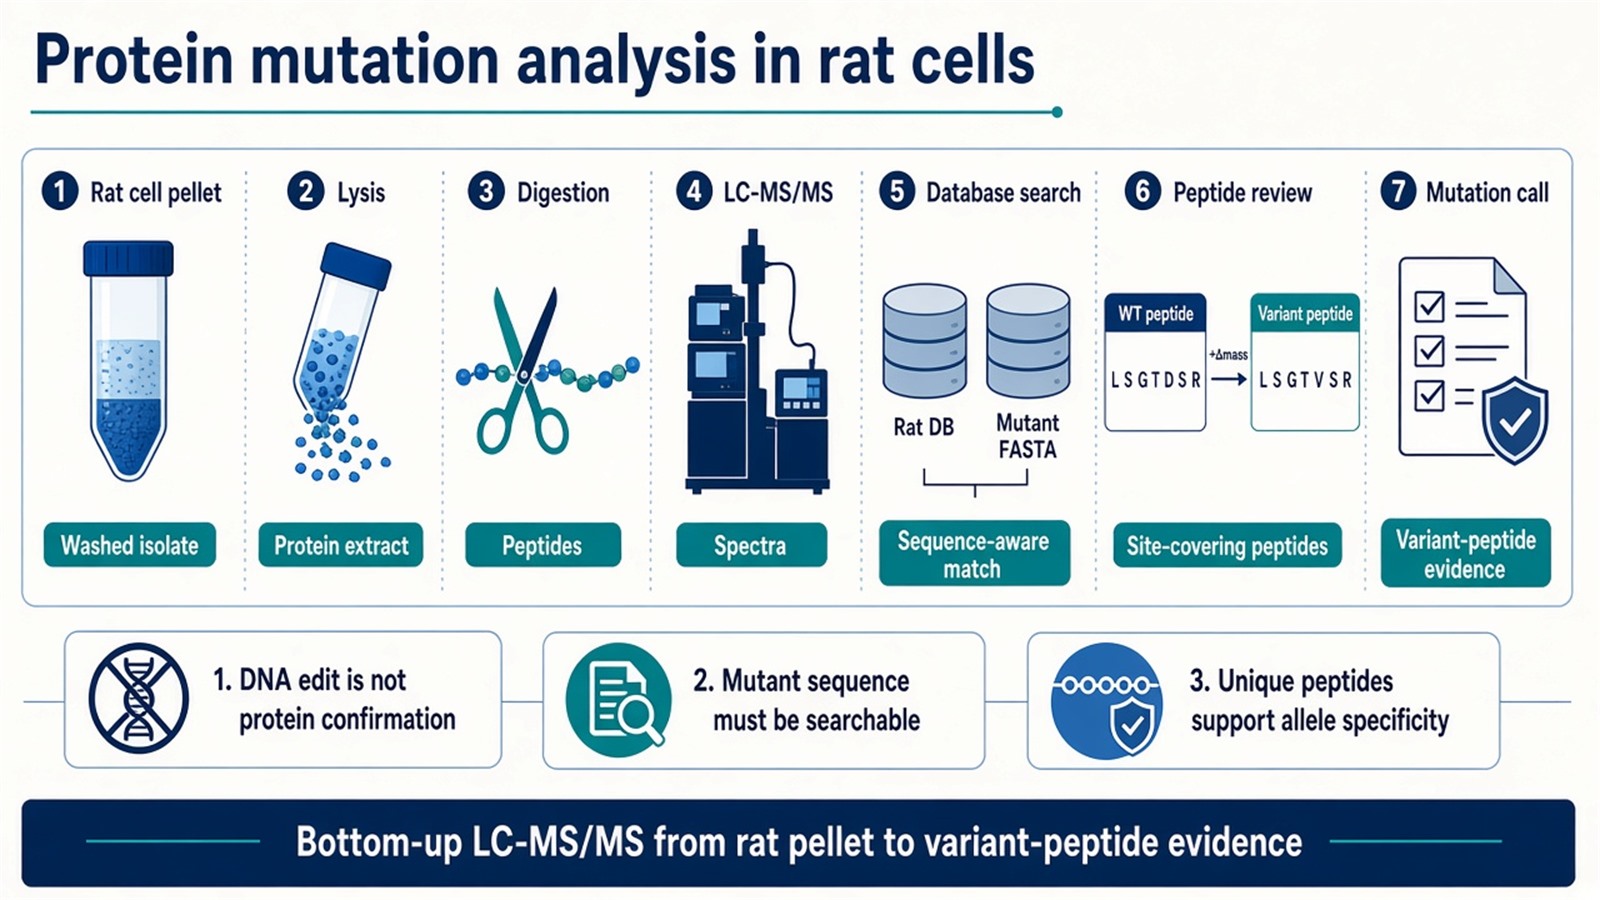
Figure 1. Protein mutation analysis in rat cells follows a pellet-to-variant-peptide order; the search database must include the expected substitution when the mutant is engineered.
Peptide-level sequence logic for known and unknown constructs is reviewed in Peptide Sequence Analysis: Methods, Workflow, and Applications in Protein Research.
Factors That Control Whether a Substitution Is Detectable
Detectability is not guaranteed by the existence of a DNA mutation. Peptide length, missed cleavages, abundance, ionization, and uniqueness all gate the result.
|
Factor |
What it changes |
What to record with the rat pellet |
|---|---|---|
|
Species database |
Whether rat accessions are searchable |
Rattus norvegicus confirmed |
|
Mutant sequence entry |
Whether the variant peptide can be scored |
Expected AA change and isoform |
|
Digestion enzyme |
Which peptides cover the mutated site |
Trypsin or multi-enzyme plan |
|
Sequence coverage around the site |
Chance of observing the substituted peptide |
Protein target and site position |
|
Unique versus shared peptides |
Allele specificity of the match |
Nearby homologs or isoforms |
|
Abundance and enrichment |
Signal for low-level variant peptides |
Whole-cell digest vs enriched target |
|
FDR or targeted QC rules |
Acceptance of the variant PSM |
Discovery search vs PRM/MRM |
A substitution that creates or destroys a trypsin cleavage site changes the peptide map itself. The wild-type and mutant digests then produce different peptide boundaries, so both forms must be considered in silico. Low-abundance mutant proteins in a mixed cell population may fall below discovery detection even when the DNA assay reports heterozygosity.
Immunoprecipitation or other enrichment can raise the chance of seeing a named target before LC-MS/MS. Enrichment still needs a sequence-aware search; it does not convert a Western blot into a variant-peptide confirmation by itself.
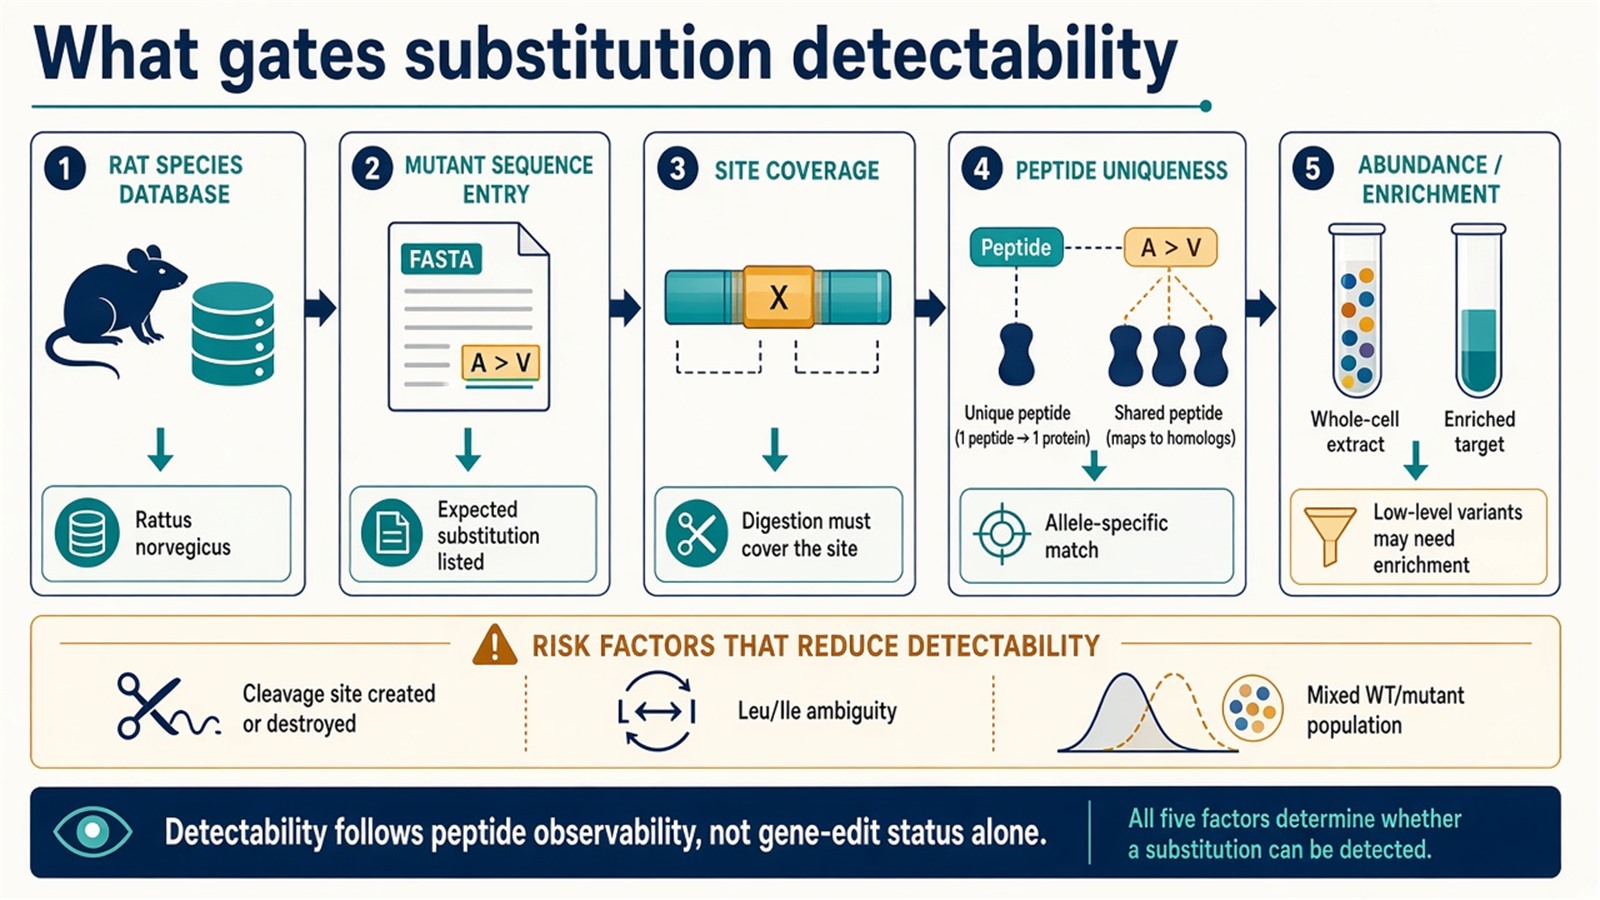
Figure 2. Variant detection depends on database design and peptide observability, not on gene-edit status alone.
How to Judge the Right Analytical Design
|
Scientific need |
Design that fits |
Do not treat as |
|---|---|---|
|
Confirm a known AA substitution in rat cells |
Custom sequence search or targeted PRM/MRM on the variant peptide |
Whole-proteome depth alone |
|
Screen an engineered mutant line for protein-level expression of the edit |
Rat DB plus mutant FASTA in discovery LC-MS/MS |
Proof that every coding SNP is expressed |
|
Map unexpected substitutions on a purified protein/peptide |
Multi-enzyme peptide mapping / mutation analysis workflow |
Untargeted whole-cell mutation census |
|
Compare abundance of WT vs mutant protein forms |
Quantitative design with both peptide forms measurable |
DNA allele frequency |
|
Assess mutation impact on interactions |
Separate IP-MS or functional assay after sequence confirmation |
Variant peptide ID as pathway proof |
If the mutation site is known and critical, prioritize peptide coverage of that site before expanding to a deep discovery list. If the site is unknown and the protein is purified, a dedicated sequence and mutation mapping route fits better than a standard cellular discovery digest. For open cellular inventories without a mutation endpoint, keep the project on ordinary rat cellular proteomics rather than forcing a mutation claim.
Researchers who already have rat cell pellets and a stated wild-type or mutant construct can also review the MtoZ Biolabs Cellular Proteomics Service for species-matched discovery planning when the first need is a protein inventory rather than a single-site confirmation.
Sample Planning for Rat Cell Mutation Projects
Rat cell lines and primary rat cells both work when species is declared and the pellet is washed. CRISPR-edited, knockdown, overexpression, and chemically treated rat cells need the same harvest discipline: remove residual medium, limit culture trypsin carryover, snap-freeze, and ship on dry ice. Mixed wild-type and mutant populations should be recorded, because a low mutant fraction reduces variant-peptide intensity.
A usable submission record names species as rat, cell type, edit or treatment, expected amino acid change if known, UniProt or construct accession, whether the target is whole-cell digest or enriched protein, and estimated cell number. Routine cellular proteomics often plans around approximately 5 × 10^6 cells as a working minimum and approximately 1 × 10^7 cells when deeper coverage is requested. Targeted confirmation of one enriched protein may use less material after purification. Those ranges are planning values, not promised detection of every substitution.
Provide the mutant protein sequence whenever the edit is known. Without that sequence, a standard rat database search cannot score the variant peptide. Contaminant databases remain useful for keratin and trypsin labeling so they are not mistaken for unexpected rat variants.
Interpretation Limits of Mutation Calls from Rat Cell MS
Accepted variant-peptide evidence supports that the substituted sequence was present in the digested material. It does not prove zygosity, allele fraction, or that every cell in the pellet expresses the mutant protein. It does not prove loss or gain of function. Functional impact needs biochemistry, phenotype, or interaction assays beyond the identification table.
Absence of the variant peptide is not proof that the DNA edit failed. The peptide may be too short, too long, poorly ionized, shared, or below detection. Leucine/isoleucine swaps and some near-isobaric changes may stay unresolved in standard MS/MS. PTMs on the same peptide can complicate both wild-type and variant scoring if modification settings are incomplete.
Pathway enrichment on proteins that happen to carry mutations annotates detected names. It does not demonstrate that the mutation caused the pathway change. Personalized-medicine or diagnostic cutoffs sit outside ordinary research reporting for rat cell discovery projects.
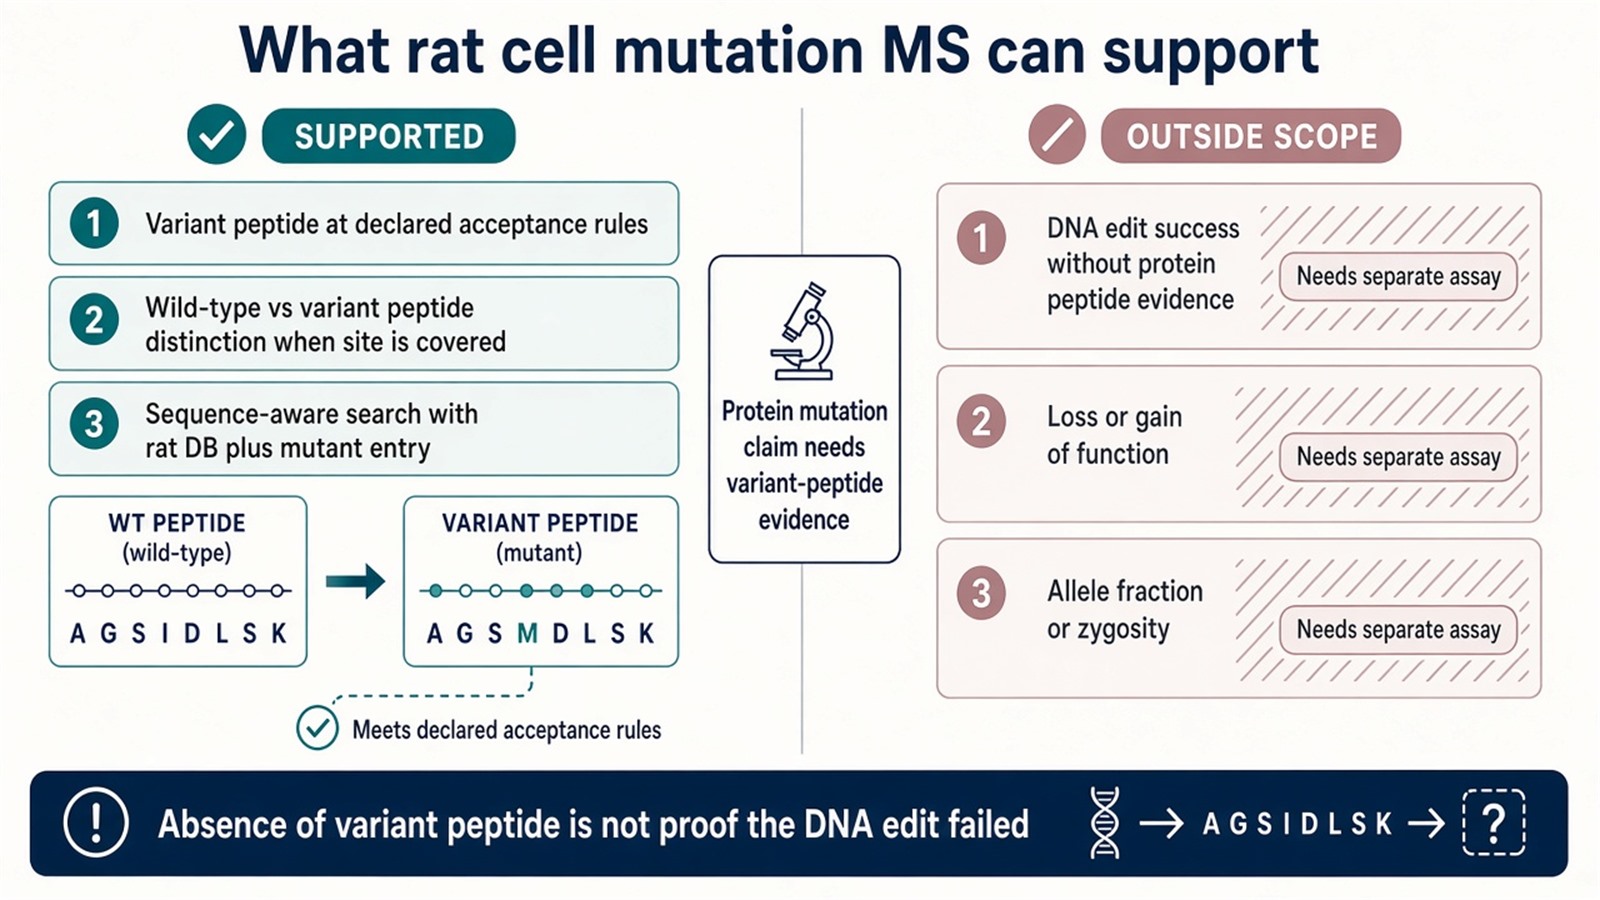
Figure 3. Protein mutation calls from rat cells rest on variant-peptide evidence; DNA status, function, and allele fraction need additional assays.
Frequently Asked Questions
1. Is CRISPR editing enough to claim a protein mutation in rat cells?
No. CRISPR changes DNA. Protein-level confirmation needs an accepted variant peptide, or another protein assay that reports the substituted product.
2. Why must the mutant sequence be added to the search database?
A standard rat proteome does not contain engineered substitutions. Without a mutant entry or variant rule, the search cannot score the altered peptide.
3. Can whole-cell discovery detect every amino acid change?
No. Detection requires a peptide that covers the site, ionizes well, and passes acceptance filters. Many coding changes remain invisible in a single trypsin digest.
4. How do protein mutations differ from PTMs in the MS table?
Mutations change the encoded amino acid sequence. PTMs add chemical groups to residues. Both shift mass, and both need explicit parameters, but they answer different biological questions.
5. When should targeted PRM or MRM be used?
Use targeted acquisition when the variant peptide sequence is already closed and needs scheduled confirmation. Keep discovery search when candidate variants are still being defined.
MtoZ Biolabs can provide analysis of protein mutations in rat cells by LC-MS/MS, including species-matched database planning, mutant-sequence search design, and review of whether discovery or targeted peptide confirmation fits the expected substitution when project samples and construct information are available.
If further evaluation of cell input, expected amino acid change, or enrichment before digestion is needed, project details can be submitted for a feasibility review.
Reference
- A.I. Nesvizhskii, R. Aebersold (2005). Interpretation of shotgun proteomic data: the protein inference problem. Mol. Cell. Proteomics, 4, 1419-1440. https://doi.org/10.1074/mcp.R500012-MCP200
- J.E. Elias, S.P. Gygi (2007). Target-decoy search strategy for increased confidence in large-scale protein identifications by mass spectrometry. Nat. Methods, 4, 207-214. https://doi.org/10.1038/nmeth1019
- Y. Zhang, B.R. Fonslow, B. Shan, M.C. Baek, J.R. Yates 3rd (2013). Protein analysis by shotgun/bottom-up proteomics. Chem. Rev., 113, 2343-2394. https://doi.org/10.1021/cr3003533
- J.R. Wisniewski, A. Zougman, N. Nagaraj, M. Mann (2009). Universal sample preparation method for proteome analysis. Nat. Methods, 6, 359-362. https://doi.org/10.1038/nmeth.1322
- B. Ma, K. Zhang, C. Hendrie, C. Liang, M. Li, A. Doherty-Kirby, G. Lajoie (2003). PEAKS: powerful software for peptide de novo sequencing by tandem mass spectrometry. Rapid Commun. Mass Spectrom., 17, 2337-2342. https://doi.org/10.1002/rcm.1196
