



An affinity pull-down assay immobilizes a known ligand on agarose or magnetic beads and asks which proteins remain after a written wash. The ligand can be a tagged protein, a small molecule, or a nucleic acid. The eluate is then read by stain, Western blot, or mass spectrometry (MS). The assay reports bead-retained association with that ligand. It does not, by itself, report an endogenous complex, a binding constant, a drug target, or a protein-interaction network. When the ligand is an antibody, the same solid-phase logic is usually named immunoprecipitation (IP) or co-immunoprecipitation (Co-IP) and is controlled with an isotype, not with an empty tag. Keeping those names separate prevents an IgG arm from being asked to serve as a GST-only lane.
Stringent washing reduces some resin binders. It also strips weak ligands. Bovine serum albumin (BSA) block can lower plastic binding on a blot and can add BSA peptides to an MS file. Neither trick converts a ligand-only control that was never processed.
Researchers who already have a tagged bait, a compound handle, or a nucleic-acid bait can also review the MtoZ Biolabs Pull-Down Assay Service for project-specific feasibility and analysis planning.
What Counts as an Affinity Pull-Down
Three ligand classes share the name and do not share the control.
A protein-tag pull-down presents GST, His, Strep, biotin, or another handle. The matched negative arm is the tag alone, or empty beads. For the GST procedure, What Is the General Procedure for GST Pull-Down Analysis of Protein–Protein Interactions? provides a complementary step list.
A small-molecule pull-down presents a compound, often through a linker on the bead, and asks which proteins from a lysate remain. The matched arm is a linker-only or inactive-analog bead. Binding to the immobilized analog is not the same as a cellular drug–target proof.
A nucleic-acid pull-down presents DNA or RNA and asks which proteins remain. Sequence-scrambled or bead-only nucleic acid is the matched arm. Sequence-specific recovery is still an in-vitro association.
Antibody beads belong with IP and Co-IP. For that capture and its isotype control, Workflow of Co-Immunoprecipitation for Protein Interaction Analysis provides a complementary antibody workflow.
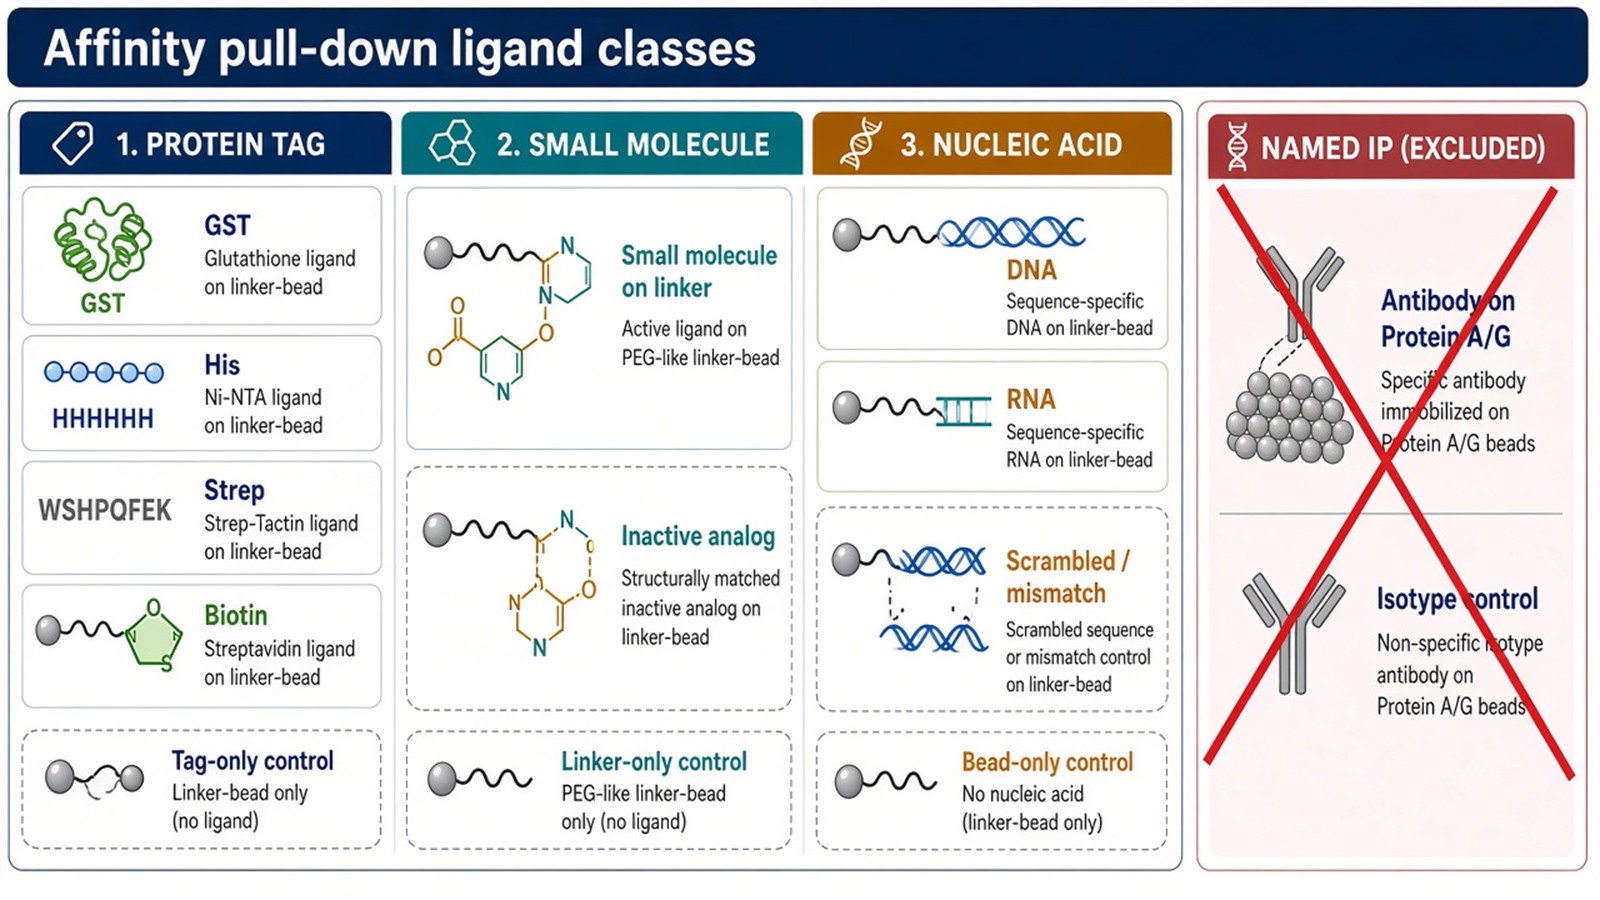
Figure 1. The shared object is a ligand on a solid phase. The control follows the ligand: tag-only, linker-only or analog, scrambled nucleic acid. Antibody capture is a different named assay.
Name the ligand and its empty-handle arm before the first binding. A field of use is not an assay design.
Typical Experimental Steps
The original sequence is kept because it is the shared chemistry.
Ligand immobilization
Couple or capture the ligand so that the binding face remains accessible. Overloading the bead raises ligand–ligand artifacts. Underloading leaves empty surface that binds abundant lysate proteins.
Target protein binding
Offer a purified protein or a clarified lysate. A purified prey is the stricter binary test. A lysate can include bridged subunits and resin binders. Time, temperature, and input mass must match the control arm.
Washing
Write salt and detergent, then keep them identical on the control. Rigorous washing is a trade-off, not a specificity guarantee.
Elution
Compete with free ligand, change pH or salt, or denature with SDS. Incomplete elution looks like a missing target. Bait and control must use the same exit. On-bead digestion is an alternative when the next readout is MS.
Identification
Stain shows protein mass. Western blot tests a nominated chain. MS opens peptide-supported protein groups. None of those detectors assigns a network or a therapeutic target. For how to read the blot or the table, How to Interpret the Results of a Pull-Down Experiment provides a complementary interpretation order.
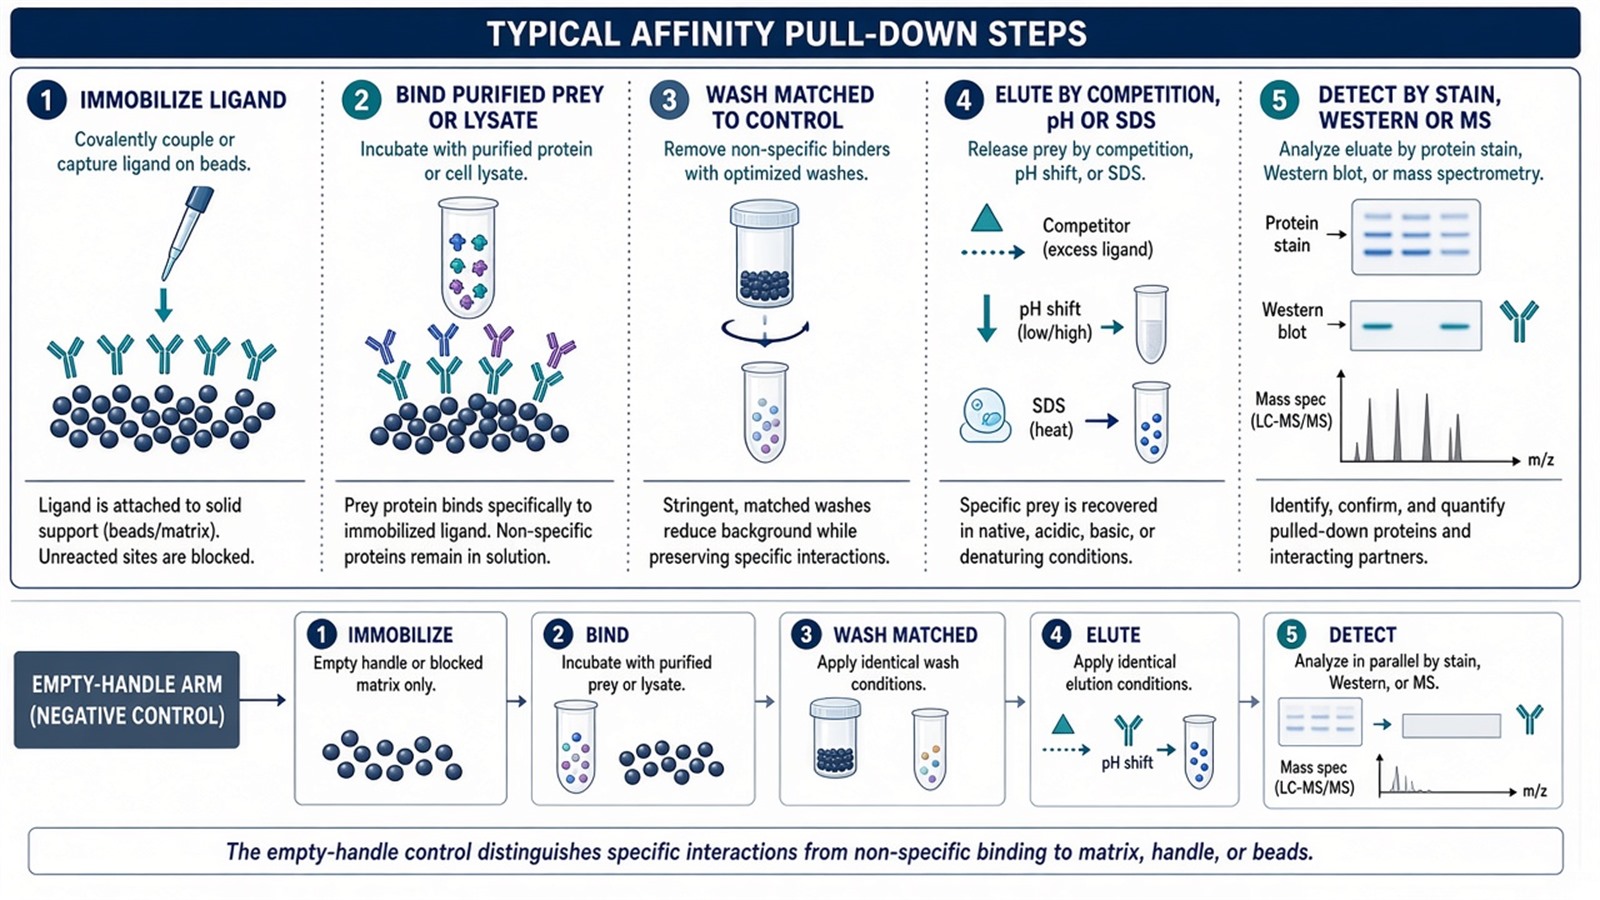
Figure 2. Five steps are common to tag, compound, and nucleic-acid pull-downs. The control ligand is prepared in the same immobilization chemistry.
What Applications the Assay Can Support
A signaling study can use a tagged pathway protein as bait and ask which wash-stable neighbors remain. That is a partner inventory around one ligand, not an intracellular regulatory map.
A compound study can ask which lysate proteins remain on an immobilized analog. That is an in-vitro binder list. Cellular target engagement needs a follow-up.
A nucleic-acid study can ask which proteins remain on a defined sequence. Chromatin or RNP language still needs the cellular context that the bead did not preserve.
Virus-host or tumor lysates can be the input. They do not change what the eluate is allowed to claim. For GST-fusion work as one ligand class, GST Pull Down Assay Service provides a complementary service path.
Limits That Stay After a Careful Wash
Nonspecific binding to the resin, the linker, or the tag is part of the method. Low-abundance partners are missed when occupancy on the ligand is low. SILAC or TMT can pair two states of what was captured. They do not raise the amount of a scarce protein that never bound the bead. Bioinformatics can annotate a shortlist. It cannot convert a linker binder into a pathway node.
BSA and detergent are optional handling tools. They are not a substitute for the empty-ligand arm.
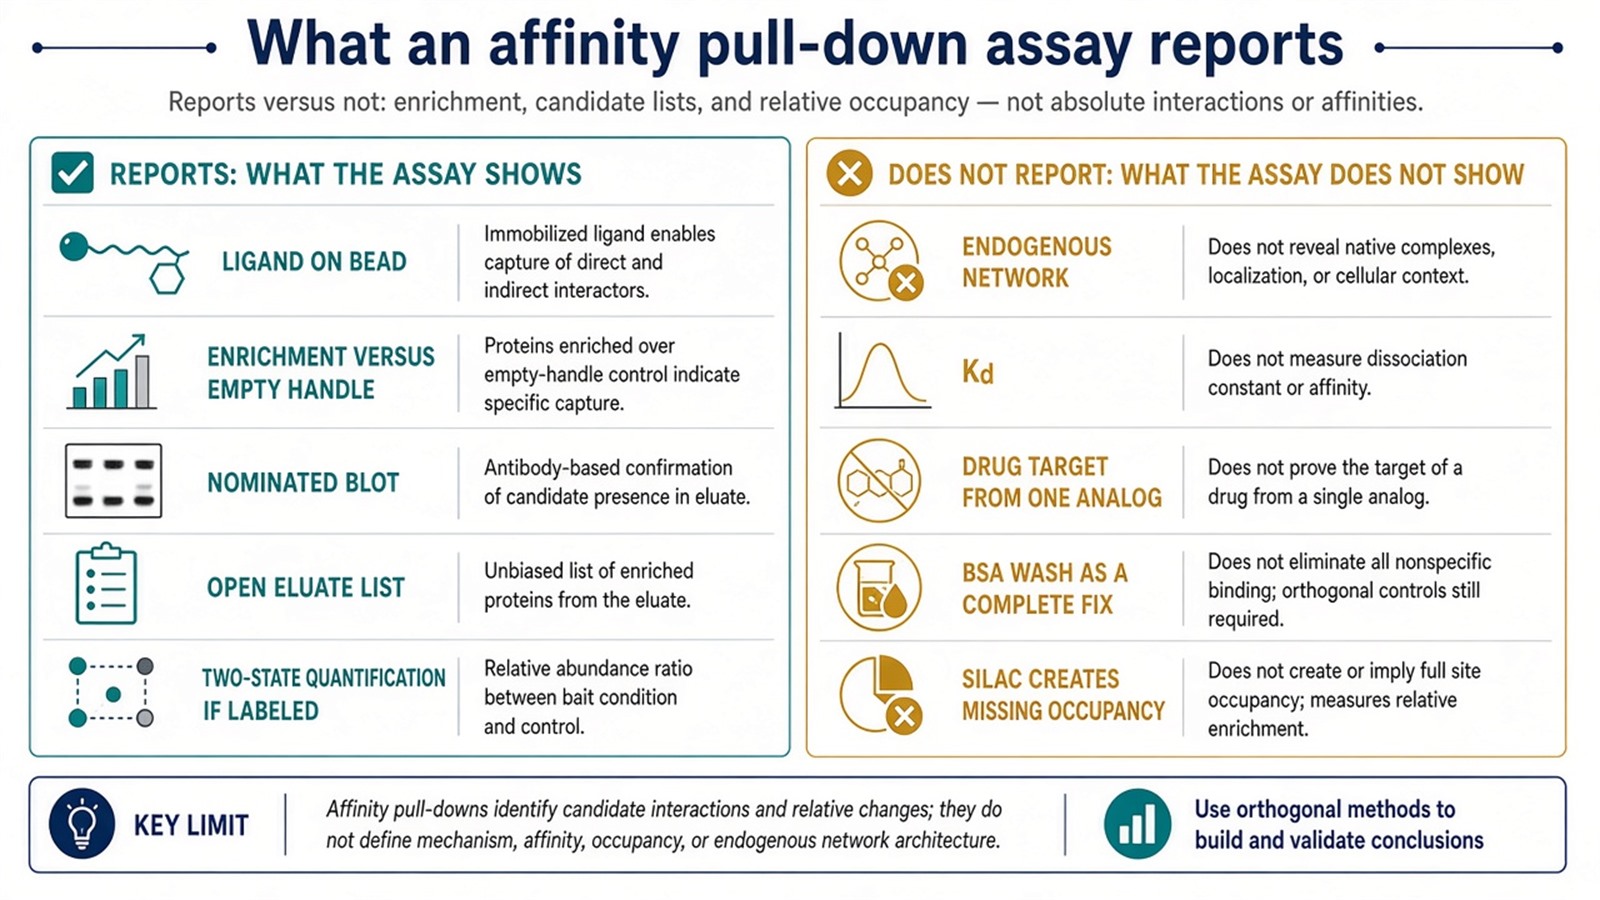
Figure 3. Supported: ligand on the bead, enrichment versus the empty handle, a nominated blot or an open eluate list. Not supported: a reconstructed interactome, a binding constant, or target nomination from one analog bead.
Experimental Limits
The assay reduces a mixture to what that ligand and wash can hold. Diagnostic language sits outside ordinary research reporting unless a separately validated assay framework is in place.
Project Planning and Service Scope
A usable affinity pull-down plan names the ligand class, the empty-handle arm, the sample (purified or lysate), the wash, the elution, and whether detection is a nominated Western or an open MS table, before the first immobilization. MtoZ Biolabs scopes tag, and related solid-phase, pull-downs with Western or Orbitrap-class LC-MS/MS. Interpretability still follows the ligand-matched control, not the instrument name.
Once the ligand is specified and the readout is an open list, the project can be scoped as pull-down MS. Researchers who need LC-MS/MS on an affinity eluate can review the MtoZ Biolabs Pull Down based Protein Analysis Service with Mass Spectrometry for project-specific sample evaluation, feasibility assessment, and workflow planning.
Frequently Asked Questions
1. What is an affinity pull-down assay?
A known ligand is immobilized on beads, a sample is bound and washed, and retained proteins are eluted and detected. The result is ligand-dependent recovery.
2. Is an antibody pull-down the same assay?
The solid-phase idea is shared. Antibody capture is named IP or Co-IP and uses an isotype control. Tag, compound, and nucleic-acid pull-downs use an empty-handle control.
3. Does a pull-down identify drug targets?
It can list proteins that remain on an immobilized compound. Cellular target status needs an orthogonal engagement assay.
4. Do SILAC or TMT fix low-abundance misses?
They quantify captured proteins across labeled states. They do not create occupancy that the ligand never provided.
5. How does this page differ from a GST protocol or a results guide?
This page defines the assay class and the ligand-matched control. GST steps and blot-or-table reading are separate jobs.
Reference
- Phizicky E.M., Fields S. (1995). Protein-protein interactions: methods for detection and analysis. Microbiol. Rev., 59, 94-123. https://doi.org/10.1128/mr.59.1.94-123.1995
- Vikis H.G., Guan K.L. (2004). Glutathione-S-transferase-fusion based assays for studying protein-protein interactions. Methods Mol. Biol., 261, 175-186. https://doi.org/10.1385/1-59259-762-9:175
- Berggård T., Linse S., James P. (2007). Methods for the detection and analysis of protein-protein interactions. Proteomics, 7, 2833-2842. https://doi.org/10.1002/pmic.200700131
- Rix U., Superti-Furga G. (2009). Target profiling of small molecules by chemical proteomics. Nat. Chem. Biol., 5, 616-624. https://doi.org/10.1038/nchembio.216
- Mellacheruvu D., Wright Z., Couzens A.L., et al. (2013). The CRAPome: a contaminant repository for affinity purification-mass spectrometry data. Nat. Methods, 10, 730-736. https://doi.org/10.1038/nmeth.2557
