



SWATH mass spectrometry here means windowed data-independent acquisition (DIA) on a liquid chromatography-tandem mass spectrometry (LC-MS/MS) system: precursors inside fixed m/z windows are fragmented in sequence while digests elute from the column, and quantification later uses extracted fragment traces under false discovery rate (FDR) control (Gillet et al. 2012). SWATH is an acquisition schedule on the mass spec, not a labeling chemistry. Matched peptide loads and stable chromatography across files still decide whether SWATH signals support a predefined relative contrast. The guide below integrates SWATH mass spec from injection through extraction handoff. It is not a vendor tuning manual, not a disease application catalog, and not a repeat of full program-level SWATH proteomics planning.
Label-free digests usually enter SWATH as separate injections. Window width, cycle time, and gradient length must fit the cohort before files accumulate. Extraction software, library provenance, and missing-value rules belong in the same methods record as the acquisition plan (Rosenberger et al. 2017).
In MtoZ Biolabs offerings, SWATH-style windowed DIA is commonly run on label-free digests after pretreatment QC, with extraction documented once window plans and run order are locked.
What SWATH Mass Spec Adds to a DIA Quantitative Project
Three mass-spec layers are often mixed in proposals and should stay separated.
Chromatography delivers peptides to the ion source with reproducible retention across the cohort. Unstable gradients or unequal loads break alignment before windows matter.
DIA SWATH acquisition fragments ions inside declared m/z windows each cycle. The plan must balance window count, isolation width, and cycle time against elution density.
Targeted extraction or library-free search turns stored MS/MS traces into peptide and protein intensities. Acquisition alone does not produce a contrast-tested matrix (Demichev et al. 2020).
SWATH mass spec project value is reproducible windowed MS/MS on matched injections plus declared extraction, not full-scan marketing language without contrast design.
Broader program framing appears in SWATH Proteomics. Pretreatment before the first SWATH injection is covered in SWATH MS Analysis and SWATH-MS Label-Free Protein Quantification Technology Analysis.
How SWATH Acquisition Connects Prep, LC, and Data Extraction
SWATH mass spec sits in the middle of the quantitative spine.
Upstream, digests must be desalted and loaded so peak capacity and ionization stay comparable. Prep mismatches appear as intensity offsets that normalization cannot always remove (Wisniewski et al. 2009).
At acquisition, a locked window table and cycle design apply to every file in the contrast. Changing windows mid cohort splits the matrix into incompatible segments.
Downstream, library-based or library-free extractors need provenance matched to species, digestion, and gradient family. OpenSWATH-style workflows are one library-based path (Quantitative Analysis of SWATH Proteomics Using OpenSWATH: A Complete Workflow Overview). General FDR and matrix QC appear in DIA Data Analysis Methods in Proteomics.
Relative discovery is the default readout. Amount on named peptides needs stable isotope-labeled internal standard (SIS) targeted assays, not open SWATH peak areas alone (Gerber et al. 2003).
Workflow From SWATH-Ready Injection to Quantitative Matrix
A practical SWATH mass spec spine usually runs as follows.
- Confirm contrast, replicate map, and relative claim.
- Verify injection-ready digests and pilot LC-MS when the matrix is new.
- Lock window plan, cycle time, and gradient method for the cohort.
- Acquire SWATH MS/MS with interleaved run order when file count allows.
- Convert raw data to analysis formats with shared conversion settings when required.
- Extract peptides under unified FDR and interference filters.
- Normalize, roll up proteins, apply missing-value policy, test predefined contrast only.
- Optional parallel reaction monitoring (PRM) on a short list after QC.
Steps 1 through 2 are prep and LC readiness. Steps 3 through 4 are SWATH mass spec proper. Steps 5 through 7 are extraction and statistics.
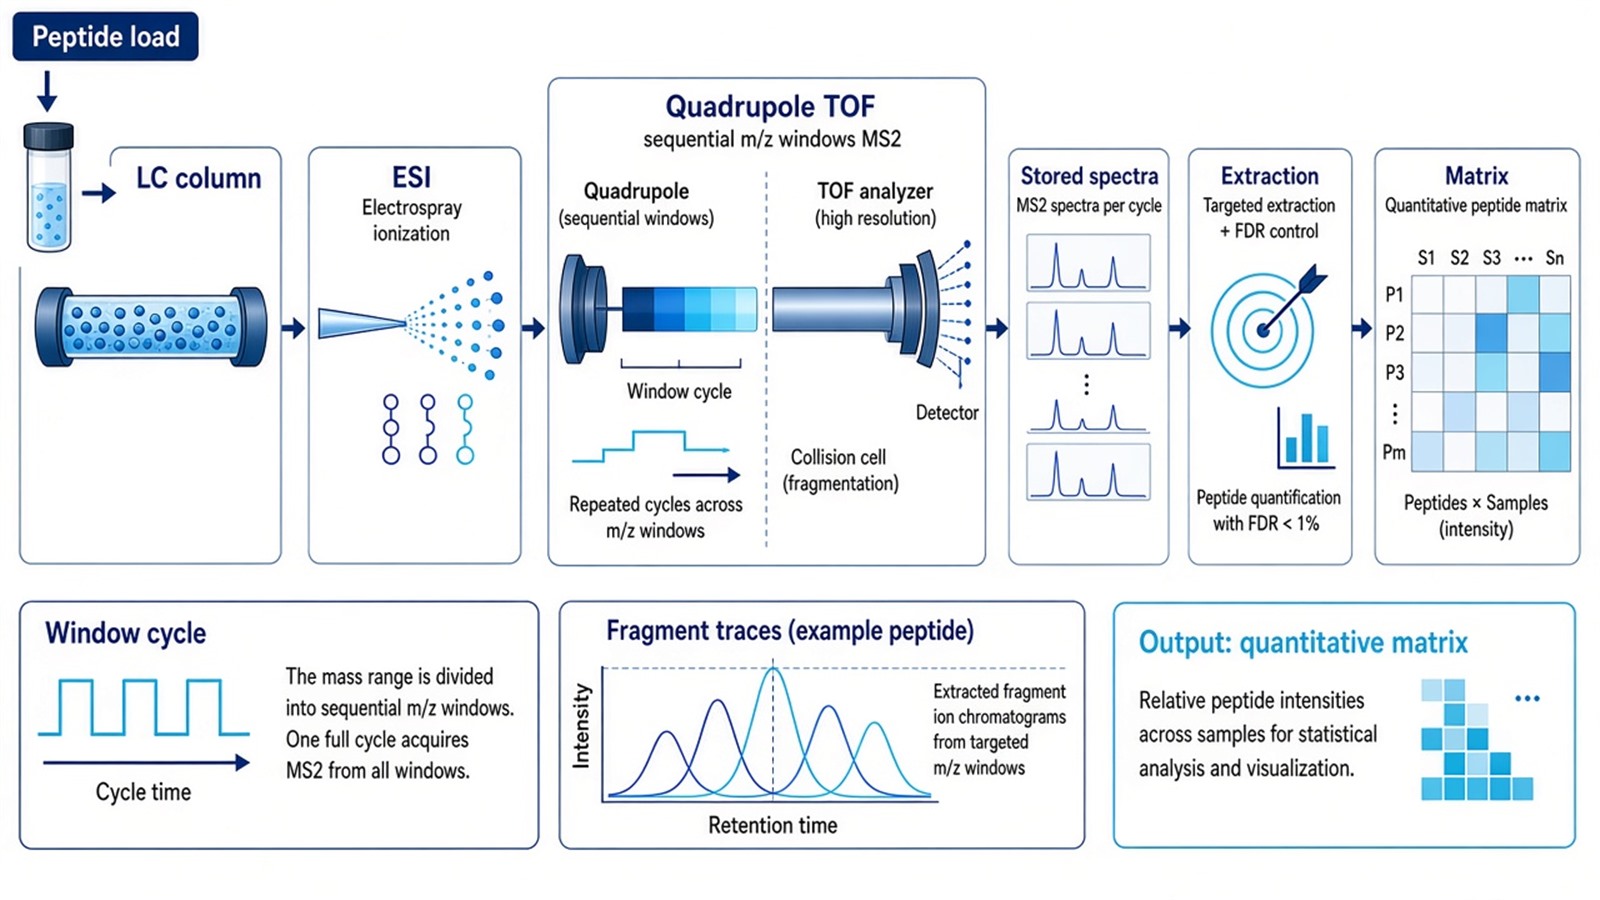
Figure 1. SWATH mass spec spans locked window acquisition and declared extraction on matched LC injections.
High-file cohort acquisition discipline aligns with High-throughput SWATH-MS Quantitative Proteomics for Biomarker Discovery. Service scoping for MtoZ projects appears in DIA based Protein Quantitative Service at MtoZ Biolabs.
Key Parameters and Limits on the Mass Spec Path
Parameter |
Role in SWATH mass spec |
Failure mode if ignored |
|---|---|---|
Peptide load per injection |
Stabilizes ion counts and peak shape |
Saturation or weak traces |
Gradient and column state |
Sets elution density versus cycle time |
Window under-coverage |
Window width and count |
Sets m/z coverage per cycle |
Gaps or excessive overlap |
Cycle time versus peak width |
Affects sampling of eluting peptides |
Distorted peak groups |
Locked plan across cohort |
Keeps extraction valid |
Mixed acquisition segments |
Run-order interleaving |
Separates batch from biology |
Injection day tracks group |
Library or library-free extractor |
Defines quantifiable peptides |
Species or digestion mismatch |
Unified FDR across files |
Controls error rate |
Per-run threshold drift |
Limits follow the same table. SWATH mass spec does not detect proteins absent from the search space or library. Low-abundance peptides can remain missing when load, ion suppression, or interference limit traces (Lazar et al. 2016). Retrospective reanalysis can mine stored spectra but still needs a declared extractor and contrast. Multi-omics overlays interpret contrasts. They do not replace matched prep or frozen group labels.
Mobility-enhanced DIA adds another separation axis before MS/MS. Labeling rules for 4D schedules appear in Is Labeling Required for 4D-DIA Quantitative Proteomics. Three-axis chemistry and route choice appear in Protein Quantification Method.
How to Choose SWATH Mass Spec for the Next Project
Use list status, chromatography constraints, and follow-up goal together.
Project need |
SWATH mass spec plan that often fits |
Recheck before committing |
|---|---|---|
Open discovery, many injections |
Locked SWATH plan, label-free files |
Contrast undefined pre-acquisition |
Complex digest, dense elution |
Narrower windows or longer gradient pilot |
Default windows without LC pilot |
Staggered sample arrival |
Separate injections, shared window table |
Mid-cohort window redesign |
Library from same project DDA |
Library-based extraction after SWATH |
Mismatched digestion in library |
No project library |
Library-free extractor with FDR QC |
Forced public library on odd matrix |
Confirmation on short list |
SWATH discovery then PRM |
Open SWATH as final assay |
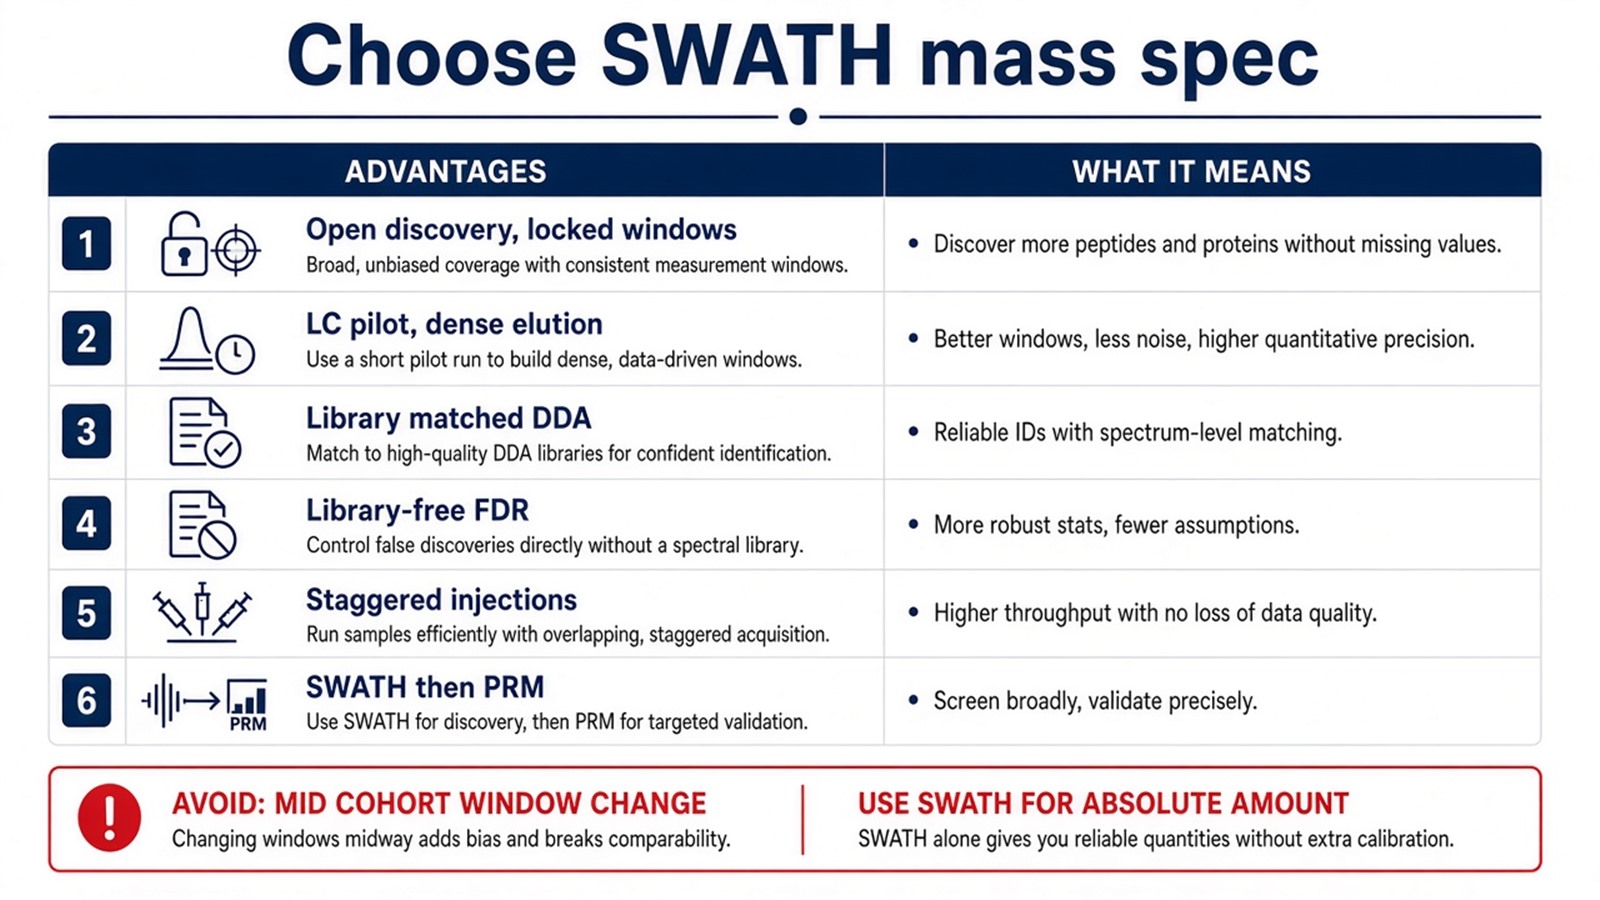
Figure 2. Choose SWATH mass spec when label-free separate injections and a locked window plan fit; PRM and SIS address closed lists and amount claims.
Application contexts for clinical, cancer, or agricultural cohorts are outlined in Typical Applications of DIA-Based Quantitative Proteomics in Clinical Research, DIA Proteomics Services for Cancer Biomarker Discovery, and Applications and Breakthroughs of SWATH Proteomics in Agricultural Research. Each cohort still needs its own prep and acquisition record.
Teams that already have matrix, contrast, and window constraints can review the MtoZ Biolabs Label-Free Quantitative Proteomics Service for SWATH mass spec planning.
Frequently Asked Questions
1. Is SWATH mass spec different from DIA?
SWATH is a common windowed implementation of DIA. Project records should still name window plans and extractors explicitly.
2. Does SWATH mass spec provide absolute protein amounts?
Not by default. Open SWATH yields relative intensities for predefined contrasts. Amount claims need SIS or another declared targeted design.
3. Must every SWATH study use a spectral library?
No. Library-free extraction is valid when documented. Library-based paths need provenance matched to the samples.
4. Can window settings change after some samples are acquired?
Changing windows mid cohort splits comparability. New settings normally require a new acquisition block and analysis plan.
5. Does SWATH mass spec remove batch effects alone?
No. Interleaved run order, prep matching, and covariates in analysis still matter. Acquisition mode does not replace design.
MtoZ Biolabs can support SWATH mass spec planning when sample type, contrast, LC constraints, and intended relative claim are documented.
If further evaluation of window piloting, library strategy, extractor choice, or discovery-to-PRM staging is needed, project details can be submitted for a feasibility review.
Reference
- S.A. Gerber, J. Rush, O. Stemman, M.W. Kirschner, S.P. Gygi (2003). Absolute quantification of proteins and phosphoproteins from cell lysates by tandem MS. Proc. Natl. Acad. Sci. U.S.A., 100, 6940-6945. https://doi.org/10.1073/pnas.0832254100
- L.C. Gillet, P. Navarro, S. Tate, H. Rost, N. Selevsek, L. Reiter, R. Bonner, R. Aebersold (2012). Targeted data extraction of the MS/MS spectra generated by data-independent acquisition: a new concept for consistent and accurate proteome analysis. Mol. Cell. Proteomics, 11, O111.016717. https://doi.org/10.1074/mcp.O111.016717
- V. Demichev, C. Messner, S. Vernardis, K.S. Lilley, M. Ralser (2020). DIA-NN: neural networks and interference correction enable deep proteome coverage in high throughput. Nat. Methods, 17, 41-44. https://doi.org/10.1038/s41592-019-0638-x
- C. Lazar, L. Gatto, M. Ferro, C. Bruley, T. Burger (2016). Accounting for the multiple natures of missing values in label-free quantitative proteomics data sets to compare imputation strategies. J. Proteome Res., 15, 1116-1125. https://doi.org/10.1021/acs.jproteome.5b00981
- G. Rosenberger, S. Liu, J. Röst, H.L. Ludwig, C. Buil, B. Borchers, R. Aebersold, L. Collins (2017). Statistical control of peptide and protein error rates in large-scale targeted data-independent acquisition analyses. Nat. Commun., 8, 14224. https://doi.org/10.1038/ncomms14224
- J.R. Wisniewski, A. Zougman, N. Nagaraj, M. Mann (2009). Universal sample preparation method for proteome analysis. Nat. Methods, 6, 359-362. https://doi.org/10.1038/nmeth.1322
