



Data-independent acquisition (DIA) quantitative proteomics gains advantages from several related techniques, not from one brand name. Shared strengths are scheduled-window fragment coverage, often fewer stochastic missing values than data-dependent acquisition (DDA), and later re-extraction of stored fragment chromatograms. Shared disadvantages are chimeric MS2 interference, software and false discovery rate (FDR) dependence, and relative-only intensities unless matched standards are added. SWATH-MS is one sequential-window, often library-based DIA route (Gillet et al. 2012). Direct or library-free DIA removes an empirical-library block but still needs interference QC (Demichev et al. 2020). Tandem mass tag (TMT) labeling is not a DIA technique. Choose among DIA routes by library logistics and cohort claim, not by a universal ranking of accuracy.
Researchers who already have defined samples, comparison groups, or a specific project objective can also review the MtoZ Biolabs Deep DIA Proteomics Service and Label-Free Quantitative Proteomics Service for project-specific feasibility and analysis planning.
Diverse DIA Techniques and Their Trade-Offs
DIA is the acquisition class: precursors in scheduled m/z windows are fragmented each cycle, and peptide quantities usually come from fragment-ion chromatograms. Within that class, common technique choices differ mainly in window scheme and extraction strategy.
Technique route |
Main advantage |
Matching disadvantage |
|---|---|---|
Shared DIA acquisition |
Fixed schedule can improve relative cohort completeness |
Window multiplexing mixes fragments |
SWATH-MS (library-based) |
Targeted library extraction supports consistent reanalysis when the library matches |
Peptides absent from the library stay invisible |
Direct / library-free DIA |
Removes separate DDA library construction as a bottleneck |
Heavier search assumptions, compute, and FDR review |
DDA library then DIA or SWATH |
Cleaner MS2 for library building, then cohort DIA completeness |
Two-stage cost; unmatched libraries transfer bias |
Gas-phase or ion-mobility aided DIA variants |
Can sharpen precursor separation in some platforms |
Still DIA-class interference and relative-only claims |
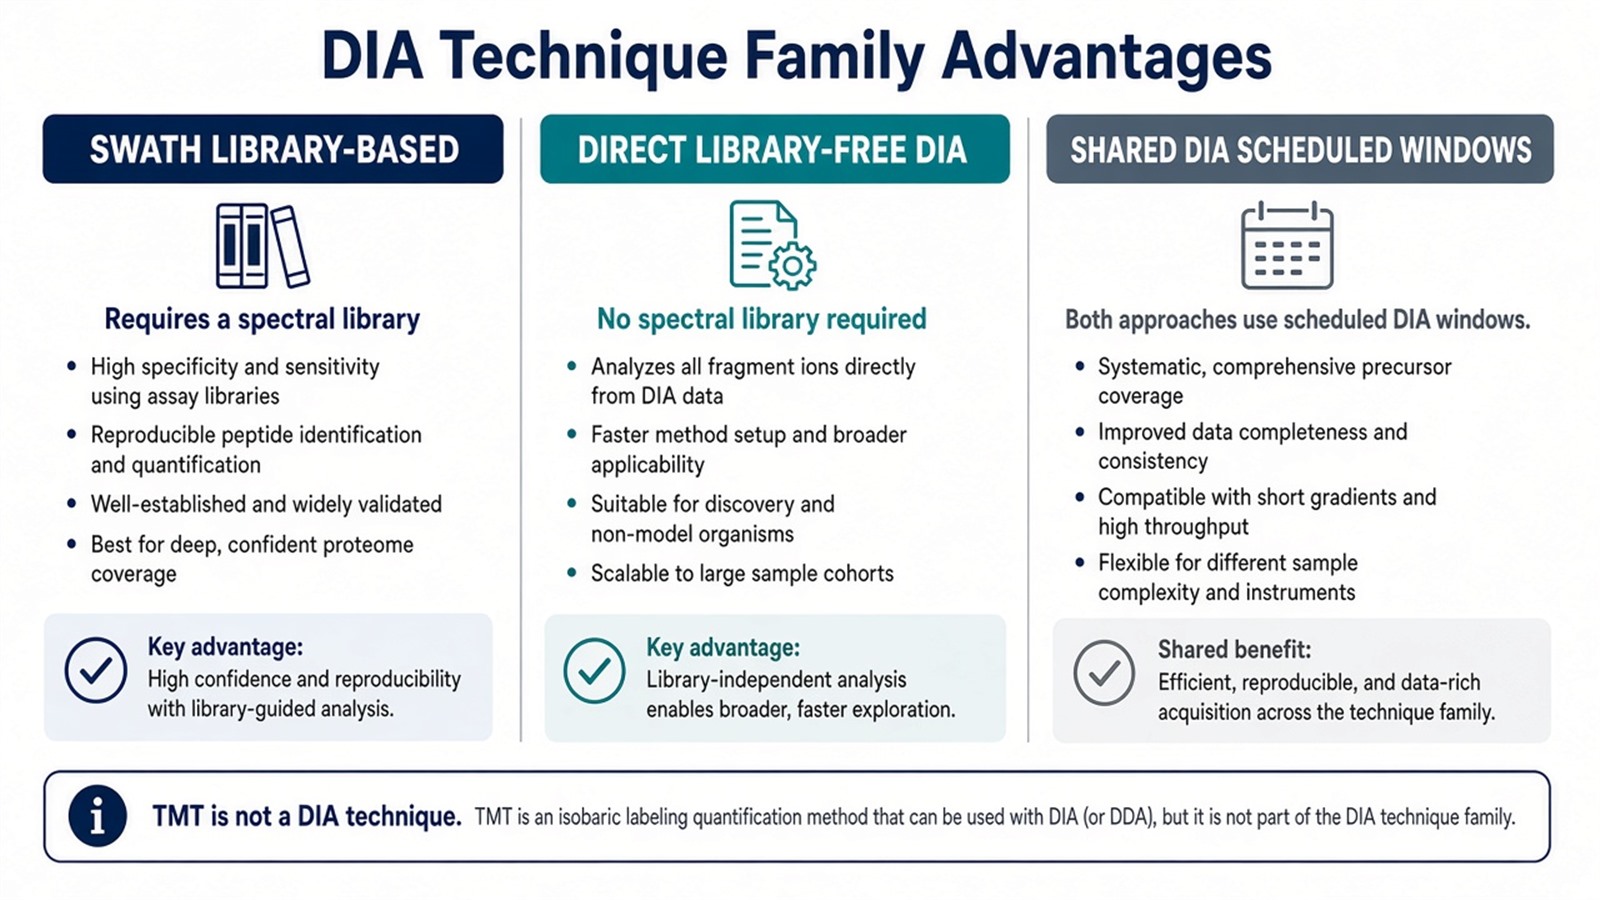
Figure 1. Diverse DIA techniques share scheduled-window acquisition; they differ in library dependence and analysis cost, and TMT is outside this family.
Technique diversity inside DIA is about extraction and window design. It is not proof that every variant is more accurate than DDA or absolute by default.
Advantages and Disadvantages of the Main Routes
SWATH-style library-based DIA
Advantage: sequential windows plus a spectral library can deliver repeatable fragment extraction across large unlabeled cohorts when the library is sample-matched. Disadvantage: library gaps and unmatched tissue or batch provenance become quantitative blind spots.
Direct or library-free DIA
Advantage: projects can start DIA quantification without a separate empirical DDA library campaign. Disadvantage: predicted or in-run libraries still need strict FDR, interference filters, and QC; compute load is often higher.
Combined DDA-then-DIA path
Advantage: DDA builds clearer spectra for a matched library, then DIA or SWATH quantifies the cohort with scheduled-window completeness. Disadvantage: two acquisition stages and the risk of using a library that no longer matches later samples.
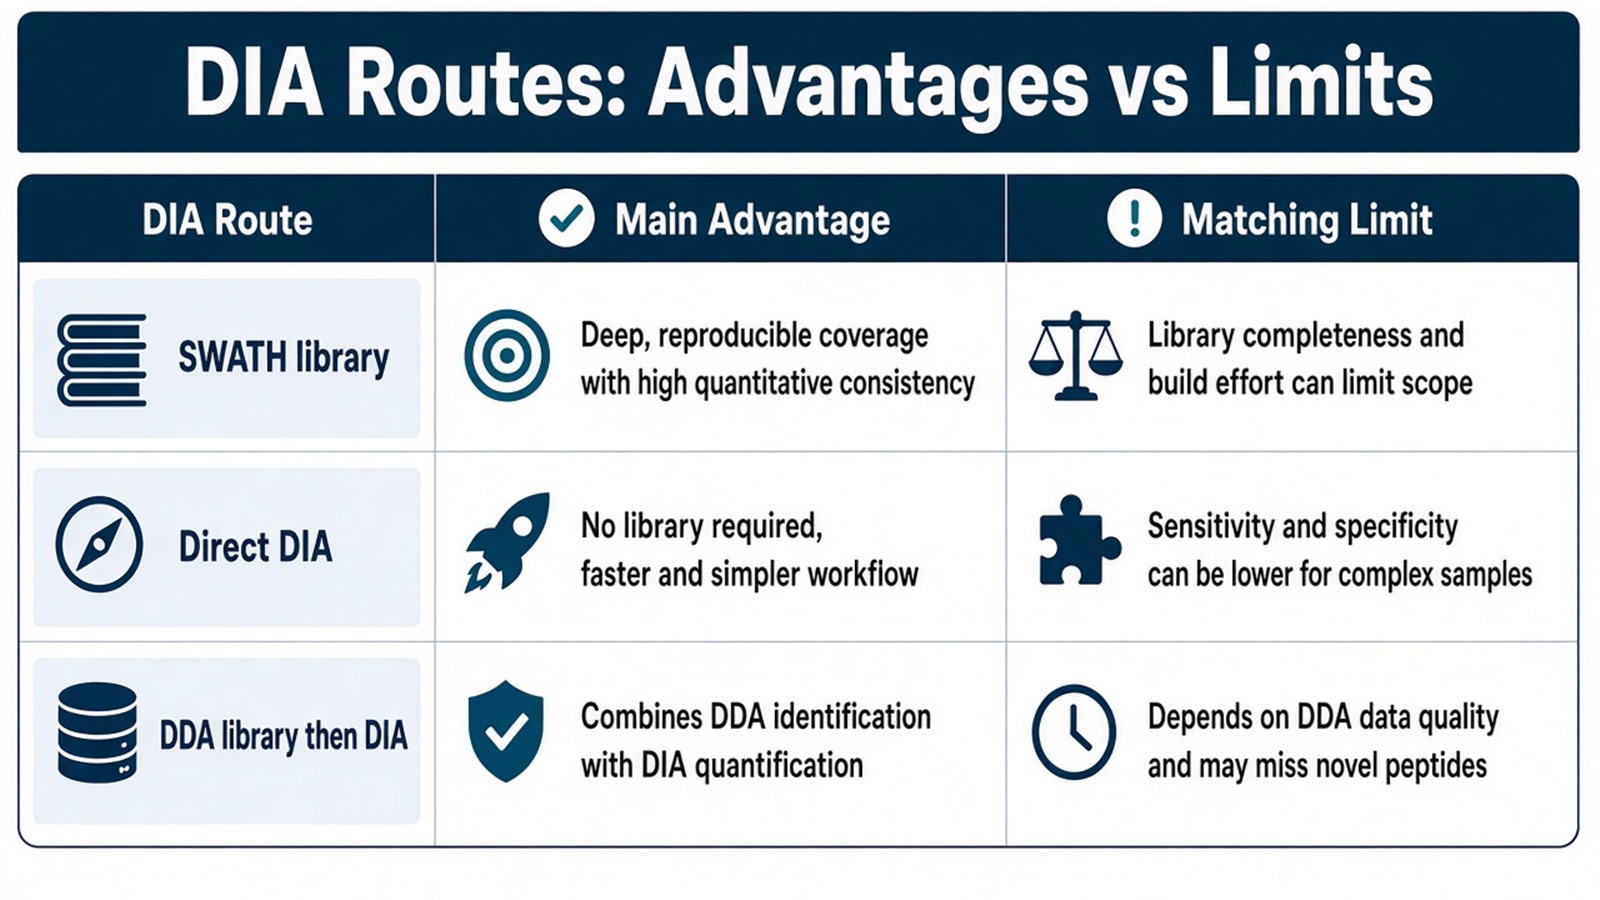
Figure 2. Each DIA route trades a different bottleneck: library coverage, library-free search cost, or two-stage design effort.
For definitions of DIA versus SWATH, see What Is DIA and SWATH? Understanding Data-Independent Acquisition in Proteomics? and What Is SWATH Mass Spectrometry. For library-free detail, see Direct DIA: High-Efficiency Proteomics without a Spectral Library. For general DIA pros and cons, see Advantages and Disadvantages of DIA in Protein Quantification and Strengths and Limitations of DIA in Protein Quantification.
Judgment Criteria: Choosing Among DIA Techniques
Study need |
Often prefer |
Reassess |
|---|---|---|
Matched empirical library already available |
SWATH-style library extraction |
Library unmatched to matrix |
Library construction is the bottleneck |
Direct / library-free DIA |
No FDR or interference QC plan |
Need cleaner MS2 before cohort quantification |
DDA library, then DIA or SWATH |
Absolute concentration required |
Identification-first or MaxLFQ-centered files |
DDA-LFQ, not DIA |
Large unlabeled cohort completeness is scarce |
Closed peptide shortlist |
PRM or MRM |
Open discovery still needed |
Closed multiplex labeling |
TMT, not a DIA extraction choice |
Confusing labeling with acquisition |
Prefer SWATH-style DIA when library logistics are solid and relative completeness matters. Prefer Direct DIA when empirical library building blocks the project. Prefer DDA first when spectral clarity or library construction dominates. Prefer PRM or MRM when targets are closed. Prefer TMT when a frozen isobaric plex is the claim, not when the question is DIA extraction strategy.
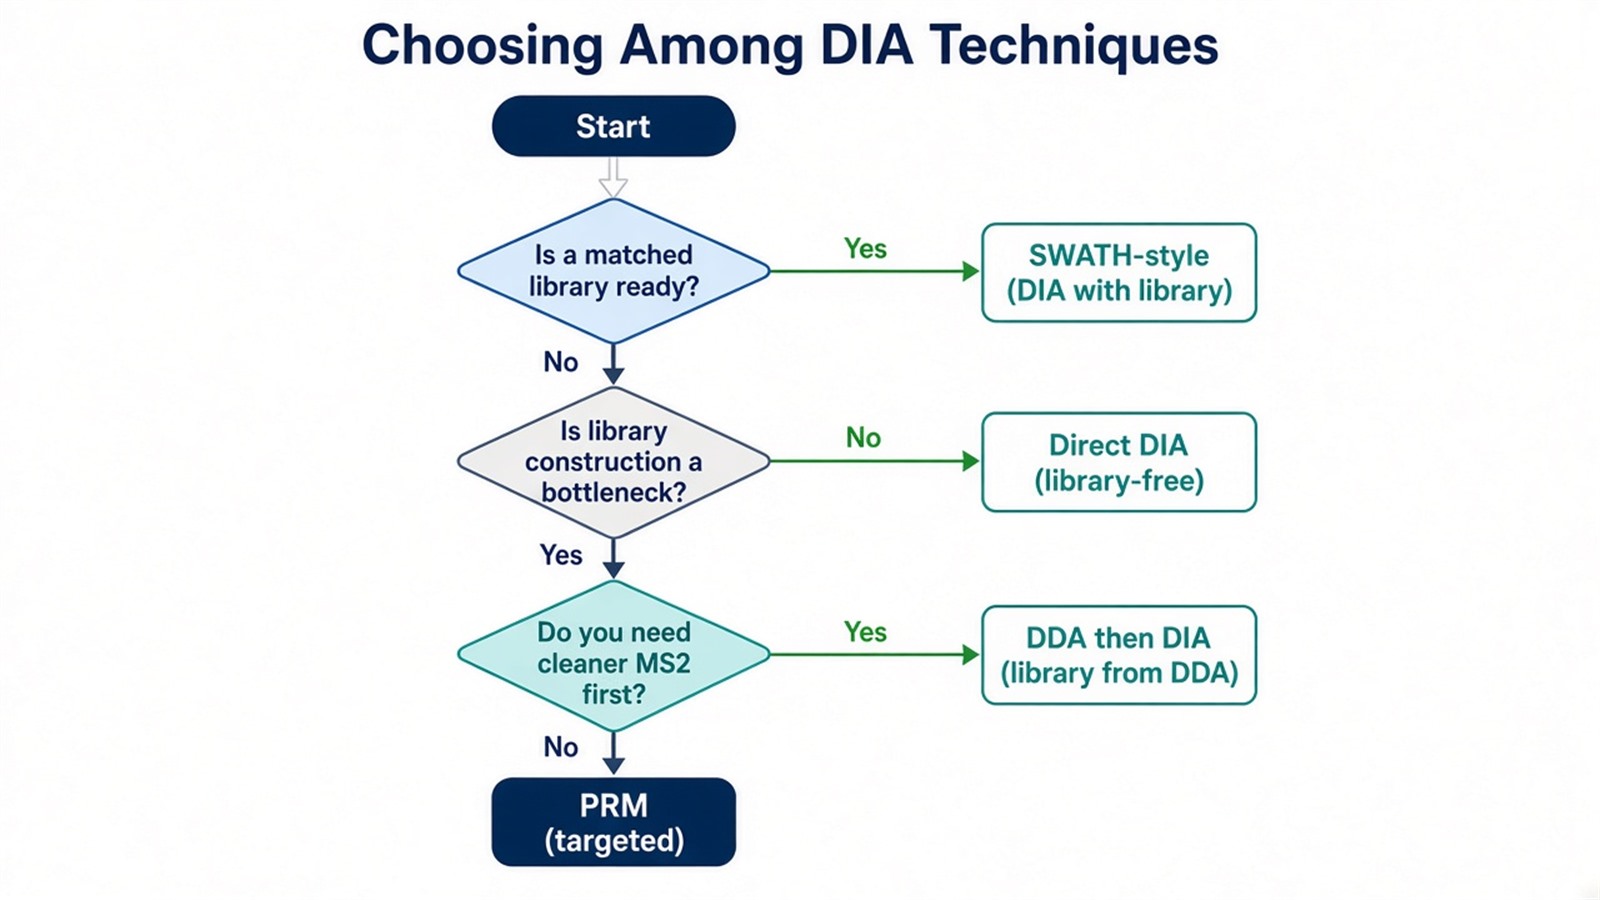
Figure 3. Select among DIA techniques from library readiness and the relative discovery claim; do not treat TMT or PRM as DIA subtypes.
For SWATH versus DDA selection, see SWATH-MS vs DDA-MS: Which Technique Is Suitable for Your Research?. For LFQ stacks, see DIA vs. DDA in Label-Free Quantitative Proteomics: A Comparative Analysis. For analysis tooling, see Essential Analysis Tools for Label-Free Proteomics: A Comprehensive Review of MaxQuant and DIA-NN.
Limits and Application Boundaries
No DIA variant removes window interference by renaming the method. Completeness is not accuracy. Ordinary DIA intensities remain relative unless stable isotope-labeled standards (SIS) and calibration are added. Ion-mobility or gas-phase fractionation variants can change precursor packing, but they stay inside the same relative claim family.
Typical applications:
- Unlabeled treatment, genotype, or time-course screens that need consistent fragment coverage.
- Complex-matrix relative discovery where stochastic DDA missingness would dominate statistics.
- Projects that reprocess stored DIA files under updated libraries or library-free searches.
- Discovery followed by PRM or MRM confirmation of a closed shortlist.
Diagnostic cutoffs and absolute stoichiometry sit outside ordinary research DIA reporting unless a separately validated assay framework is in place.
Once the DIA route, library or library-free path, and relative comparison design are defined, the project can be evaluated as a DIA quantification study. Researchers can review the MtoZ Biolabs Deep DIA Proteomics Service for sample evaluation, feasibility assessment, and workflow planning. For analysis pitfalls, see Common Pitfalls in DIA Proteomics Data Analysis and How to Avoid Them.
Frequently Asked Questions
1. Is TMT one of the diverse DIA techniques?
No. TMT is isobaric labeling with reporter-ion readout. DIA is an acquisition and extraction class for scheduled windows.
2. Is SWATH better than DIA?
SWATH-MS is a DIA implementation, usually library-based. Advantages follow library quality and window QC, not the SWATH name alone.
3. Does Direct DIA remove all DIA disadvantages?
No. It mainly removes empirical-library construction. Chimeric windows, FDR control, and relative-only claims remain.
4. Are higher throughput and higher accuracy automatic with every DIA technique?
No. Throughput and usable accuracy depend on chromatography, window design, library or search settings, and QC, not method class alone.
5. When should a project keep DDA instead of any DIA technique?
Keep DDA when identification-first spectra, MaxLFQ-centered files, or a matched DDA library for later DIA are the priority.
Reference
- L.C. Gillet, P. Navarro, S. Tate, H. Rost, N. Selevsek, L. Reiter, R. Bonner, R. Aebersold (2012). Targeted data extraction of the MS/MS spectra generated by data-independent acquisition: a new concept for consistent and accurate proteome analysis. Mol. Cell. Proteomics, 11, O111.016717. https://doi.org/10.1074/mcp.O111.016717
- V. Demichev, C.B. Messner, S.I. Vernardis, K.S. Lilley, M. Ralser (2020). DIA-NN: neural networks and interference correction enable deep proteome coverage in high throughput. Nat. Methods, 17, 41-44. https://doi.org/10.1038/s41592-019-0638-x
