



Protein quantification in biological research means measuring protein abundance under a stated claim, usually from peptide surrogates after digestion and liquid chromatography-tandem mass spectrometry (LC-MS/MS). Relative discovery methods such as label-free quantification (LFQ) and tandem mass tags (TMT) compare the same protein across samples without reporting concentration by default (Cox et al. 2014; Thompson et al. 2003). Targeted parallel or multiple reaction monitoring (PRM or MRM) confirms a closed peptide list and remains relative unless standards are added. Absolute quantification uses matched standards such as AQUA peptides and a calibration model to place named peptides on an amount scale (Gerber et al. 2003). Colorimetric bulk assays and immunoassays answer different questions and are not synonyms for MS proteome quantification.
Researchers who already have defined samples, comparison groups, or a specific project objective can also review the MtoZ Biolabs Label-Free Quantitative Proteomics Service, TMT Quantitative Proteomics Analysis Service, and Targeted Proteomics Service for project-specific feasibility and analysis planning.
Definition: What Protein Quantification Means Here
In MS-based proteomics, quantification starts from proteotypic peptides. Proteins are extracted, digested, separated, ionized, and identified from fragment evidence. Quantitative values then come from MS1 peak areas, fragment chromatograms, reporter ions, or light-to-heavy ratios, depending on the route. The defended output is same-protein relative change, closed-list confirmation, or calibrated peptide amount, not a single universal number for every protein in a cell.
Bulk colorimetric assays such as BCA or Bradford estimate total protein load for input matching. ELISA and Western blot quantify selected antigens with antibody reagents. Those tools remain useful, but they do not replace an LC-MS/MS relative matrix or an AQUA concentration table for a proteomics claim.
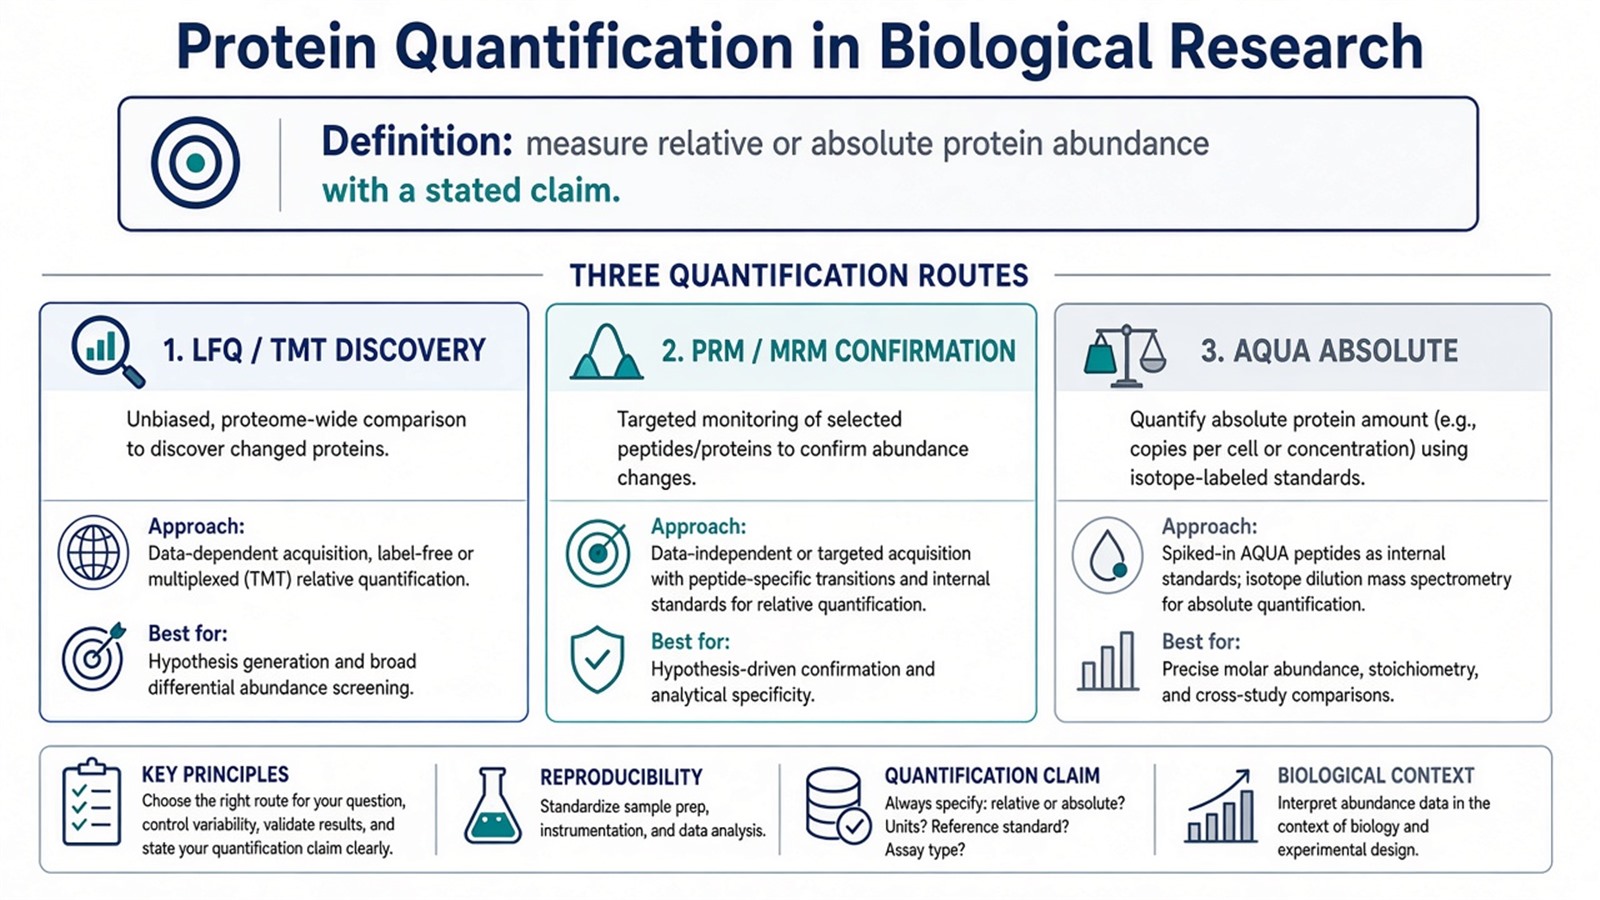
Figure 1. Protein quantification in biological research is defined by the claim and the peptide-based MS readout, not by bulk dye assays alone.
Prospects follow what the claim can defend: relative change, confirmed targets, or calibrated amount.
Principle Layers Behind Common MS Routes
Layer |
Relative discovery |
Relative targeted |
Absolute |
|---|---|---|---|
Typical methods |
LFQ (DDA or DIA), TMT / iTRAQ |
PRM or MRM without SIS |
AQUA / SIS with PRM or MRM |
Core principle |
Compare the same peptide across samples under one plan |
Schedule known peptides and integrate peak areas |
Convert light-to-heavy response through calibration |
Sample handling |
Matched digestion; independent runs or labeled plex |
Closed peptide list and QC |
Extra aliquots for calibrators and stated spike timing |
Default output |
Fold change or relative intensity |
Confirmation of nominated targets |
Amount or concentration for named peptides |
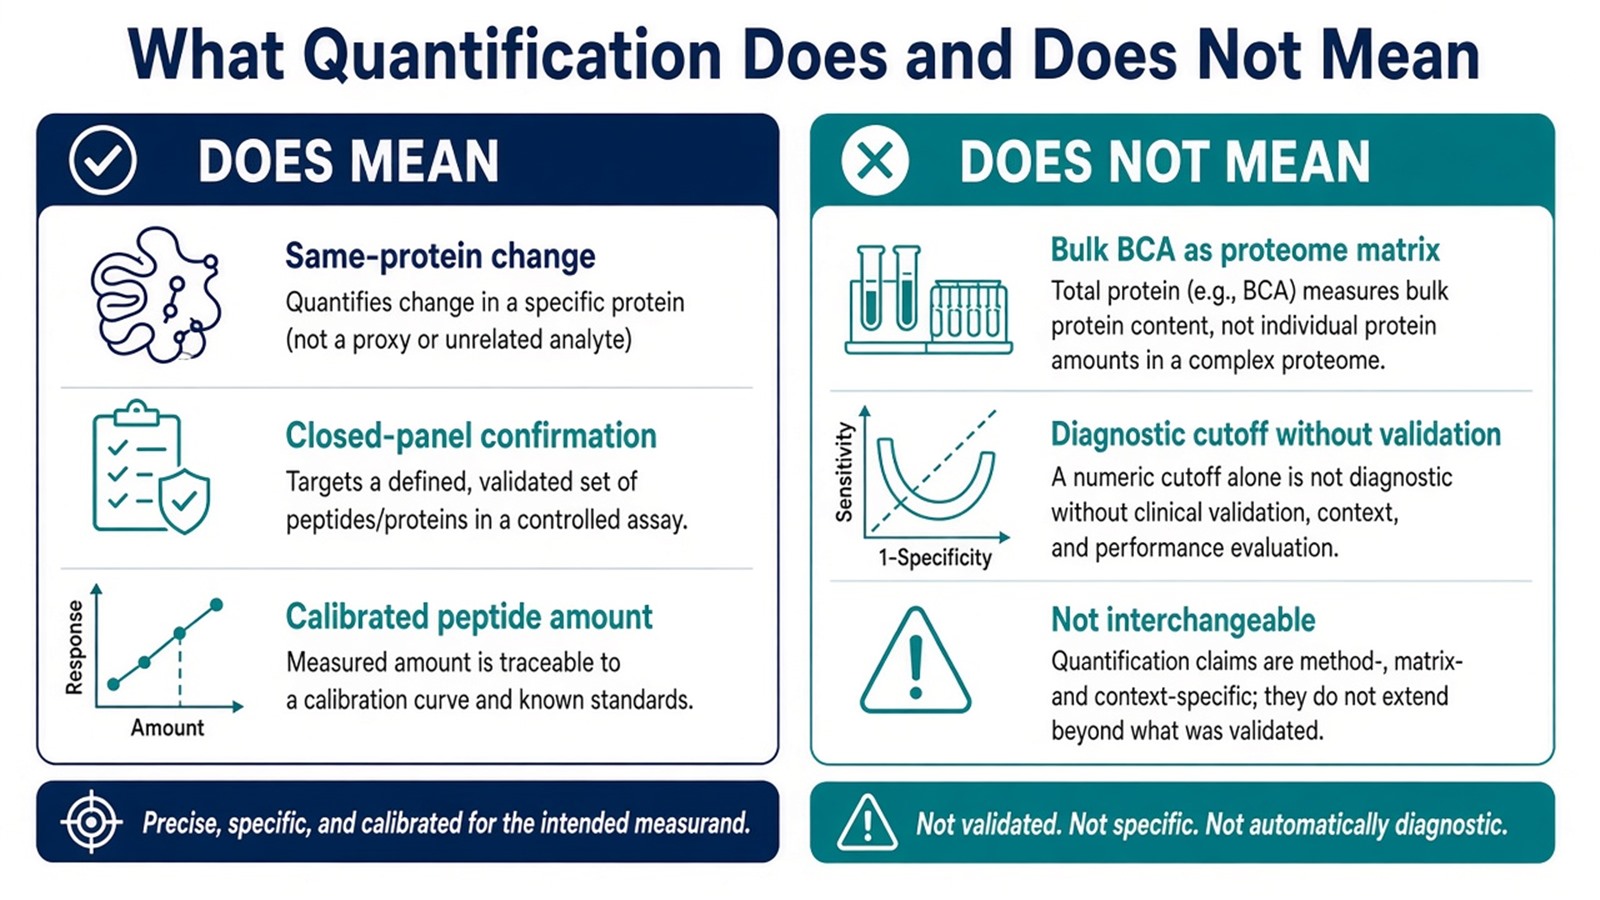
Figure 2. MS protein quantification supports relative discovery, closed-list confirmation, and calibrated amount; it does not automatically equal total-protein colorimetry or clinical cutoffs.
For a broader claim map, see Protein Quantification Measured by Mass Spectrometry. For labeled versus unlabeled families, see A Comprehensive Overview of Labeled and Label-Free Quantitative Proteomics Methods. For sample and pretreatment constraints, see Absolute and Relative Quantitative Proteomics Analysis.
Judgment Criteria: Realistic Prospects by Study Need
Biological research need |
Quantitative prospect that often fits |
Reassess |
|---|---|---|
Mechanism or treatment screens |
Relative LFQ or DIA across matched digests |
Absolute units required from day one |
Multi-arm genotype or dose panels that fit one kit |
Relative TMT reporter ratios |
Open enrollment after mixing |
Biomarker candidates already nominated |
Relative PRM or MRM confirmation |
Inventory still open |
Concentration, stoichiometry, or specification |
Absolute AQUA / SIS assay |
Fold change alone is enough |
Clinical diagnostic cutoff |
Validated assay framework beyond ordinary research reporting |
Treating any discovery table as a cutoff |
Prefer open relative discovery while the protein inventory is still changing. Prefer targeted confirmation when peptides are closed. Prefer absolute assays only when the decision sentence needs amount. Prefer immunoassay transfer after the MS claim is defined, not as a substitute for stating relative versus absolute design.
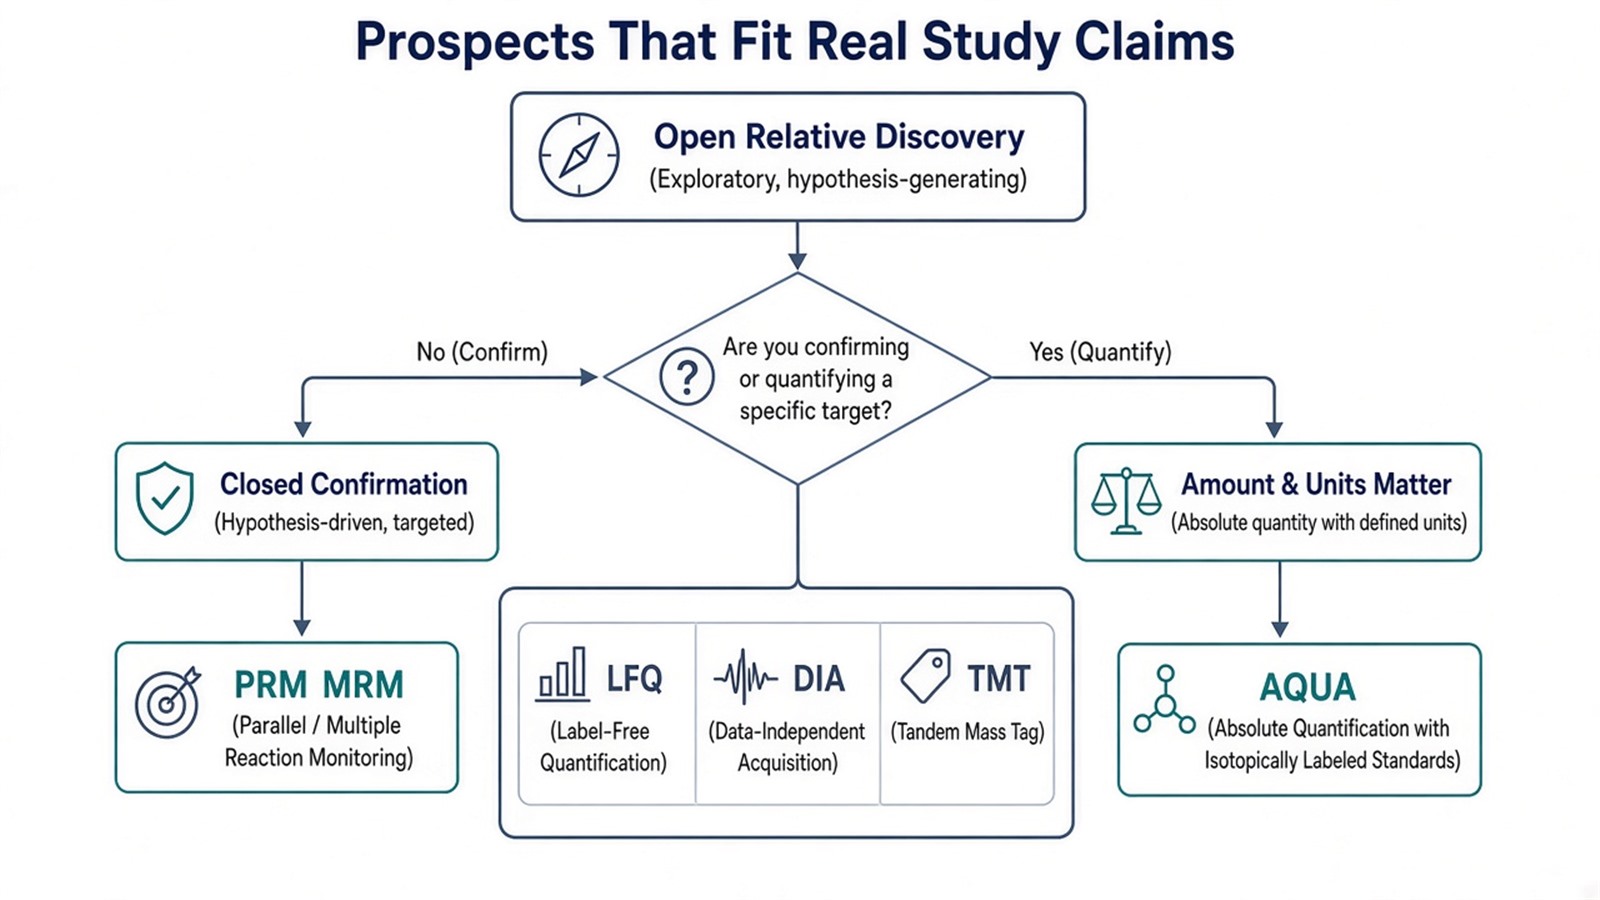
Figure 3. Application prospects are claim-dependent: discovery, confirmation, and absolute amount are different research stages, not interchangeable labels.
For LFQ versus isobaric selection, see Comparing LFQ and Isobaric Labeling Strategies in Proteomics Quantification. For DIA trade-offs, see Advantages and Disadvantages of DIA in Protein Quantification. For targeted and absolute trade-offs, see PRM and MRM in Targeted Proteomics: Advantages and Limitations and Advantages and Disadvantages of AQUA Technique.
Limits and Application Boundaries
MS intensities do not become concentration without standards and calibration. Completeness in DIA is not the same as accuracy when windows are chimeric. TMT same-run comparison does not remove ratio compression. Pathway enrichment from a differential list shows overrepresentation relative to the quantified background; it does not prove pathway activity in vivo.
Typical applications with a realistic prospect:
- Disease-mechanism and pharmacology screens that need relative protein change across defined groups.
- Biomarker discovery followed by PRM or MRM confirmation of a shortlist.
- Multi-condition panels that fit one TMT kit and need same-run relative contrast.
- Named-protein amount when process control, stoichiometry, or research validation requires units.
Diagnostic decision rules, companion-diagnostic claims, and nutritional total-protein certificates sit outside ordinary research proteomics reporting unless a separately validated assay framework is in place.
Once the claim, sample plan, and panel status are defined, the project can be scoped as relative discovery, targeted confirmation, or absolute quantification. Researchers can review the MtoZ Biolabs Deep DIA Proteomics Service when unlabeled DIA is part of the design, and the Absolute Quantitative Analysis (AQUA) Service when calibrated amount is required.
Frequently Asked Questions
1. Does protein quantification always mean absolute concentration?
No. Most biological discovery work is relative. Absolute amount needs matched standards and calibration.
2. Are BCA and MS proteomics the same prospect?
No. BCA estimates bulk protein load. MS proteomics quantifies named peptides under a stated claim.
3. Can one MS method cover every biological research need?
No. Open discovery, closed confirmation, and calibrated amount use different designs and QC.
4. What limits the near-term prospect of clinical cutoffs from discovery MS?
Ordinary research tables lack the analytical validation framework required for diagnostic decisions.
5. Where do immunoassays fit after MS quantification?
They can transfer a shortlist after peptides and claims are defined. They do not replace stating relative versus absolute MS design.
Reference
- J. Cox, M.Y. Hein, C.A. Luber, et al. (2014). Accurate proteome-wide label-free quantification by delayed normalization and maximal peptide ratio extraction, termed MaxLFQ. Mol. Cell. Proteomics, 13, 2513-2526. https://doi.org/10.1074/mcp.M113.031591
- A. Thompson, J. Schaefer, K. Kuhn, S. Kienle, J. Schwarz, G. Schmidt, T. Neumann, C. Hamon (2003). Tandem mass tags: a novel quantification strategy for comparative analysis of complex protein mixtures by MS/MS. Anal. Chem., 75, 1895-1904. https://doi.org/10.1021/ac0262560
- S.A. Gerber, J. Rush, O. Stemman, M.W. Kirschner, S.P. Gygi (2003). Absolute quantification of proteins and phosphoproteins from cell lysates by tandem MS. Proc. Natl. Acad. Sci. U.S.A., 100, 6940-6945. https://doi.org/10.1073/pnas.0832254100
