



Sample type and sample preparation decide whether absolute or relative quantitative proteomics is feasible before any mass spectrometry method is named. Relative discovery needs matched extraction, digestion, and loading across the cohort, then either unlabeled independent injections for label-free quantification (LFQ) or data-independent acquisition (DIA), or amine-compatible labeling and pooling for tandem mass tags (TMT). Absolute AQUA-style assays add a closed peptide list, matrix-matched calibrators, and a recorded spike time for heavy standards, because a peptide added after digestion does not correct protein loss during lysis (Gerber et al. 2003; Cox et al. 2014). Scarce input, residual amine buffers, leftover detergent, and unmatched loads create artifacts that later software cannot repair.
Researchers who already have defined samples, comparison groups, or a specific project objective can also review the MtoZ Biolabs Label-Free Quantitative Proteomics Service, TMT Quantitative Proteomics Analysis Service, and Absolute Quantitative Analysis (AQUA) Service for project-specific feasibility and sample planning.
Sample Constraints Before Absolute or Relative Claims
Relative quantification compares the same protein or peptide across samples that share one coherent preparation plan. Absolute quantification places a named peptide on an amount scale using matched standards and calibration. Sample chemistry and aliquot budget often close one of those claims before acquisition starts.
Sample or prep situation |
Relative claim often still open |
Absolute claim often blocked until redesigned |
|---|---|---|
Matched lysates or tissues, equal protein load |
LFQ, unlabeled DIA, or TMT if amines are free |
Still needs reserved material for calibrators and QC |
Clinical biofluids arriving over weeks |
Independent LFQ or DIA injections |
Hard to freeze a TMT plex; AQUA needs a closed panel and extra aliquots |
Scarce biopsy or low-input digest |
Unlabeled relative if labeling would waste material |
Calibration curves consume sample that may not exist |
Residual Tris, ammonium bicarbonate excess, or amine buffers |
LFQ or DIA after cleanup |
TMT labeling completeness fails; SIS peptides still need a clean digest |
FFPE or archival blocks |
Relative LFQ if digestion can be matched |
Peptide recovery may be too unstable for a concentration curve |
Targets not yet chosen |
Open relative discovery |
AQUA and PRM or MRM panels cannot be built |
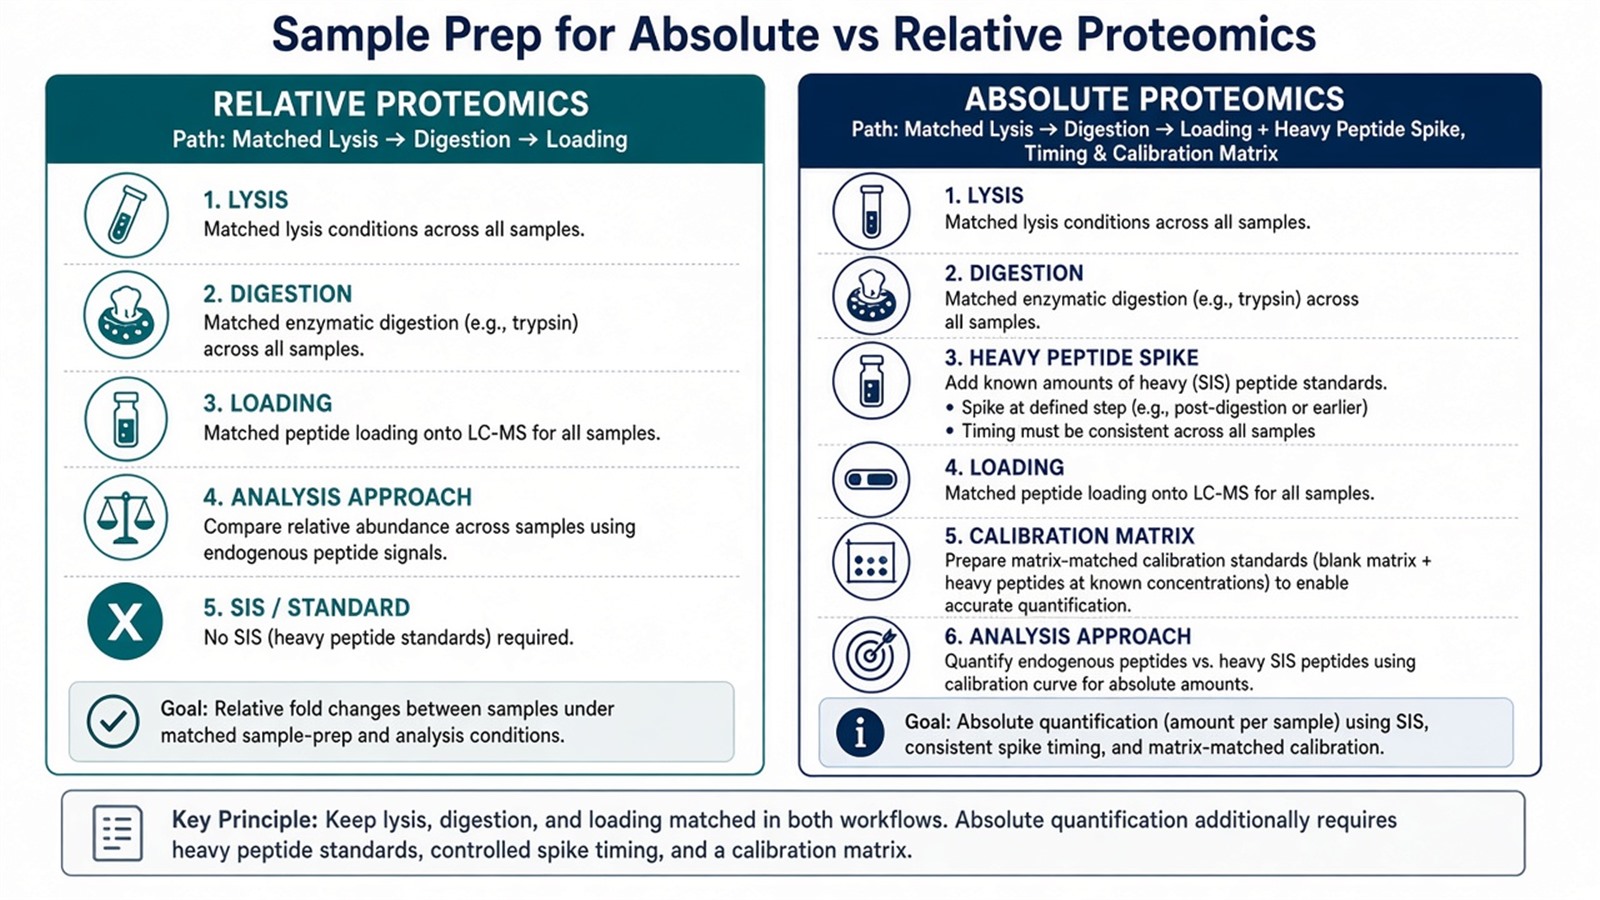
Figure 1. Absolute versus relative claims follow sample constraints first: matched prep enables relative comparison, while amount units need reserved aliquots, a closed peptide list, and defined spike timing.
Prep failures become quantitative artifacts. Software cannot reconstruct protein that was lost before the standard or the label entered the sample.
Shared Prep Path and Claim-Specific Branches
Most bottom-up projects share an early path: lyse under a declared buffer, clarify, assay protein, reduce and alkylate, digest (usually with trypsin), and clean up peptides. Loading should be matched on protein amount, not on volume alone, when groups will be compared.
After that common path, the claim branches:
- Relative unlabeled: each digest is an independent liquid chromatography-tandem mass spectrometry (LC-MS/MS) run under DDA-LFQ or DIA. Keep detergents low enough for chromatography and randomize run order.
- Relative isobaric: each digest is amine-labeled, quenched, and pooled into one TMT plex. Residual free amines and unequal loads before labeling become channel artifacts (Thompson et al. 2003).
- Absolute: a heavy AQUA or other stable isotope-labeled standard (SIS) peptide is spiked at a stated step, then acquired with the endogenous light peptide by PRM or MRM against a matrix-matched curve (Gerber et al. 2003).
Spike timing is a prep decision, not a software option. A peptide added after digestion reports peptide recovery in that digest. It does not automatically correct lysis loss or incomplete protein digestion. Protein-level or concatenated labeled standards can move the correction earlier when that is part of the claim, at higher design cost.
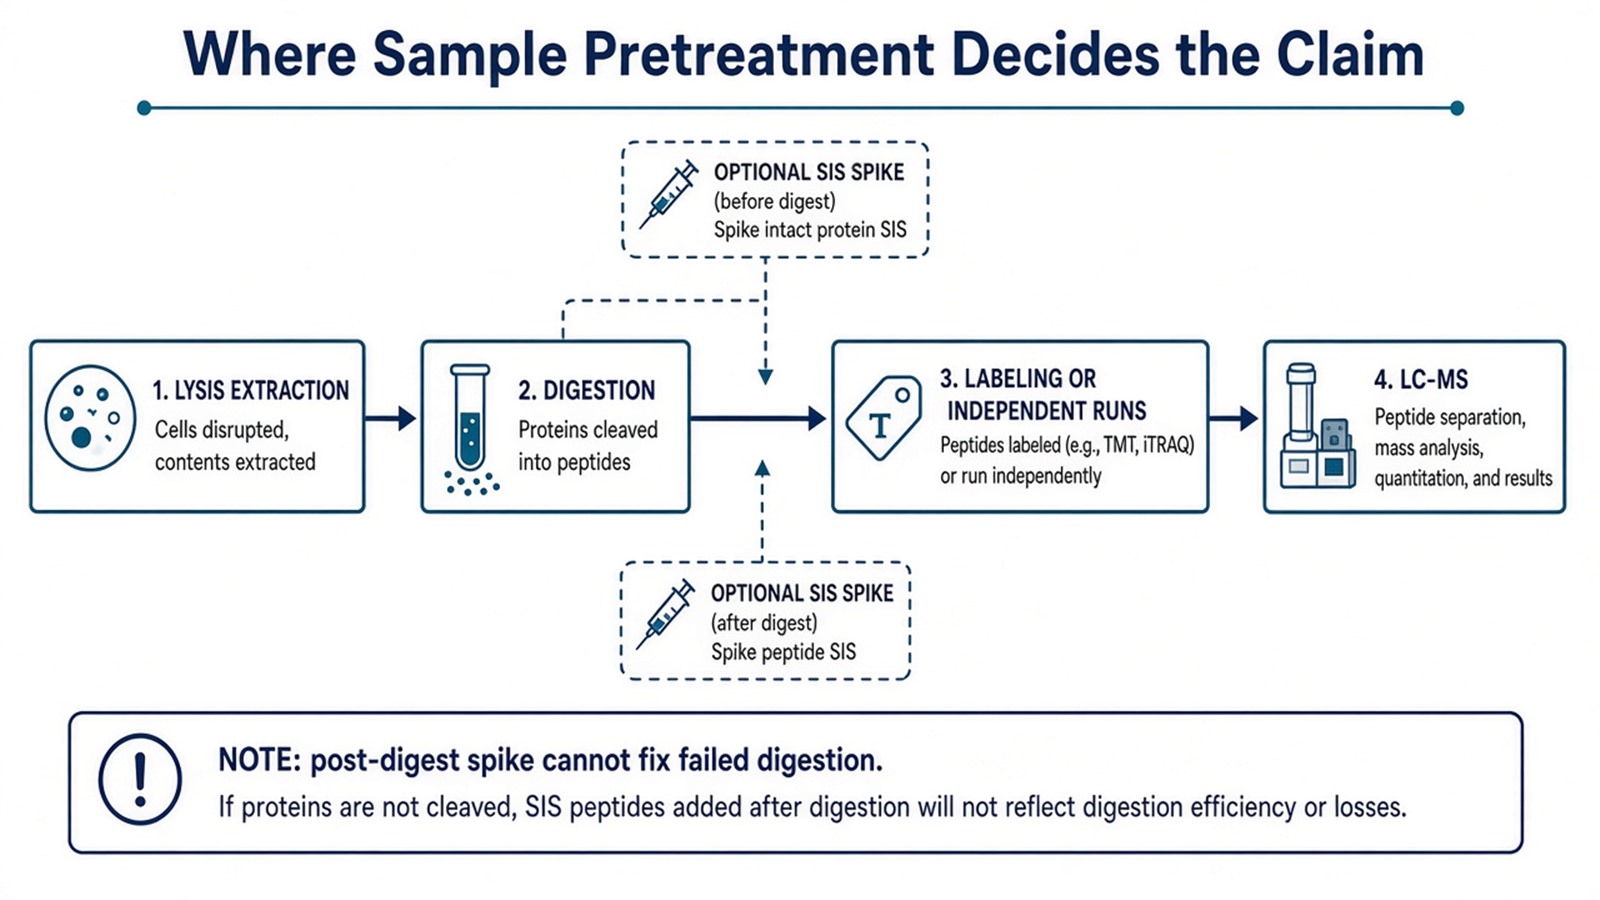
Figure 2. Absolute and relative routes share early prep; they diverge at labeling, pooling, or SIS spike timing.
For operational unlabeled prep, see What Is Label-Free Analysis? LFQ Principles, Workflow, Advantages, and Limitations. For TMT labeling-to-pool steps, see A Detailed Workflow of TMT-Based Quantitative Proteomics. For spike timing and peptide-level amount, see Absolute Quantification in Targeted Proteomics.
Judgment Criteria: Prep That Fits Each Claim
Prep question |
Prefer relative LFQ or DIA |
Prefer relative TMT |
Prefer absolute AQUA |
Reassess |
|---|---|---|---|---|
Can every sample share one lysis and digest recipe? |
Yes, independent injections |
Yes, then label and pool |
Yes, plus calibrators in similar matrix |
Protocol still drifting |
Is enrollment closed before mixing? |
No need to freeze a plex |
Required |
Required for the calibrated panel |
Samples still arriving |
Are amines free for NHS-style tags? |
Not required |
Required |
Not the AQUA bottleneck |
Buffer exchange incomplete |
Is extra material available for a curve and QC? |
Helpful but not defining |
QC pools still needed |
Required for a defended amount |
Only one aliquot remains |
Are proteotypic peptides already chosen? |
No, discovery can stay open |
No, discovery can stay open |
Yes, the list must be closed |
Inventory still changing |
Prefer relative unlabeled prep when samples arrive over time, labeling chemistry is risky, or input is too scarce to spend on tags. Prefer TMT prep when equal, amine-clean digests can be frozen in one kit. Prefer AQUA prep when amount is required, the peptide list is closed, and matrix-matched calibrators can be built without exhausting the study samples.
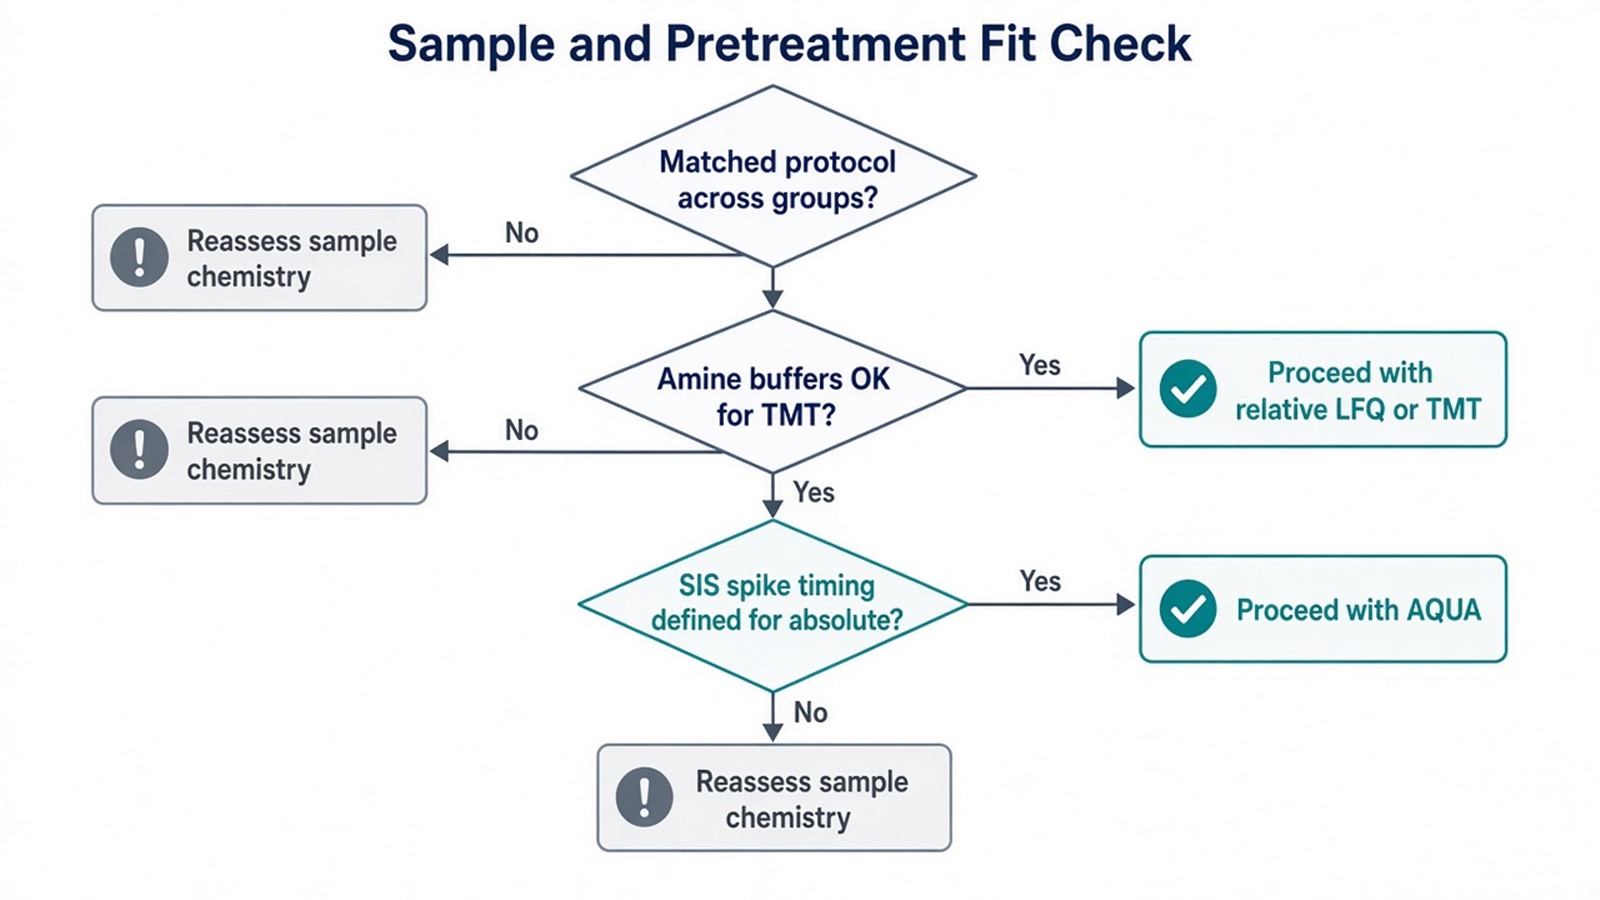
Figure 3. If buffers, aliquot budget, or enrollment rules fail, change the quantitative claim rather than forcing an incompatible prep.
For claim-level method maps beyond prep, see Protein Quantification Measured by Mass Spectrometry and Comparing LFQ and Isobaric Labeling Strategies in Proteomics Quantification. For AQUA trade-offs after prep is feasible, see Advantages and Disadvantages of AQUA Technique. For panel planning, see Targeted Proteomics Experimental Design.
Limits and Typical Sample Applications
Matched prep does not convert relative intensities into concentration. SWATH-style or other DIA acquisition remains a relative unlabeled route unless SIS and calibration are added; it is not an absolute method by name. iBAQ-style intensity proxies rank proteins within a sample under model assumptions and are not a substitute for an AQUA curve.
Typical applications tied to sample handling:
- Cell or tissue treatment panels with equal protein loads: relative LFQ, DIA, or TMT depending on enrollment and amine chemistry.
- Longitudinal plasma or serum collections: relative unlabeled discovery, with later AQUA only on a closed shortlist if reserved aliquots exist.
- Low-input biopsies: relative unlabeled if tags or calibration curves would consume the only digest.
- Process or specification samples that already have named peptides: AQUA with documented spike timing and matrix-matched calibrators.
Diagnostic cutoffs and nutritional total-protein certificates sit outside ordinary research absolute or relative proteomics reporting unless a separately validated assay framework is in place.
Once sample type, buffer compatibility, aliquot budget, and spike or labeling timing are defined, the project can be scoped as relative discovery or absolute targeted work. Researchers can review the MtoZ Biolabs Label-Free Quantitative Proteomics Service and Absolute Quantitative Analysis (AQUA) Service for sample evaluation and workflow planning.
Frequently Asked Questions
1. Can I switch from relative to absolute on the same digest without extra material?
Usually no. Absolute assays need calibrators, QC levels, and often a reserved aliquot. Plan the extra input before the only tube is digested.
2. Does adding a heavy peptide after digestion give protein concentration?
It reports peptide amount in that digest under the calibration. Lysis and digestion losses that happened earlier stay outside the correction unless the design uses an earlier protein-level standard.
3. Are TMT and AQUA both absolute because they use labels?
No. TMT reporters support relative channel ratios inside a plex. AQUA uses known standard amounts and calibration for a concentration claim.
4. Which buffers block TMT but still allow relative LFQ?
Amine-containing leftover buffers compete with TMT reagents. After cleanup, unlabeled LFQ or DIA can still proceed if digestion quality is matched.
5. Is SWATH-MS an absolute quantification prep route?
No. SWATH-style DIA is scheduled-window relative acquisition. Absolute units still need matched standards and calibration.
Reference
- S.A. Gerber, J. Rush, O. Stemman, M.W. Kirschner, S.P. Gygi (2003). Absolute quantification of proteins and phosphoproteins from cell lysates by tandem MS. Proc. Natl. Acad. Sci. U.S.A., 100, 6940-6945. https://doi.org/10.1073/pnas.0832254100
- J. Cox, M.Y. Hein, C.A. Luber, et al. (2014). Accurate proteome-wide label-free quantification by delayed normalization and maximal peptide ratio extraction, termed MaxLFQ. Mol. Cell. Proteomics, 13, 2513-2526. https://doi.org/10.1074/mcp.M113.031591
- A. Thompson, J. Schaefer, K. Kuhn, S. Kienle, J. Schwarz, G. Schmidt, T. Neumann, C. Hamon (2003). Tandem mass tags: a novel quantification strategy for comparative analysis of complex protein mixtures by MS/MS. Anal. Chem., 75, 1895-1904. https://doi.org/10.1021/ac0262560
