



Label-free quantitative proteomics compares extracted peptide or protein intensities across liquid chromatography-tandem mass spectrometry (LC-MS/MS) runs without isobaric or metabolic tags on the discovery path (Gillet et al. 2012). Four-dimensional data-independent acquisition (4D-DIA) adds ion mobility separation before windowed MS/MS, then quantifies through mobility-aware extraction under false discovery rate (FDR) control (Meier et al. 2020). The guide below integrates label-free chemistry with 4D-DIA acquisition, prep, and project fit. It is not a generic proteomics survey, not a disease diagnostic promise, and not a claim that 4D-DIA replaces every other quant route.
Label-free means separate unlabeled injections for most open cohorts. 4D-DIA names mobility-enhanced windowed DIA on those digests. Relative contrast across predefined groups remains the default claim. Amount on named peptides needs stable isotope-labeled internal standard (SIS) targeted assays, not open label-free intensities alone (Gerber et al. 2003).
In MtoZ Biolabs offerings, label-free 4D-DIA discovery is evaluated when platform scope supports mobility-aware schedules and cohort design fits separate injections with matched prep.
How Label-Free Chemistry Relates to 4D-DIA Acquisition
Three labels belong in the project brief and should stay separated.
Label-free chemistry keeps digests unlabeled through injection. Quantification compares fragment traces across files after normalization.
4D-DIA acquisition adds ion mobility and windowed MS/MS on each run. Mobility can reduce crowded MS/MS overlap in complex digests when methods are locked.
Extraction and statistics turn stored spectra into a protein matrix. FDR, interference filters, and missing-value policy belong in the same record as window and mobility settings (Rosenberger et al. 2017; Demichev et al. 2020).
Label-free 4D-DIA project value is a contrast-tested relative matrix on matched unlabeled digests, not a technology slogan without replicate design.
Tagging rules when mobility is present appear in Is Labeling Required for 4D-DIA Quantitative Proteomics. Mechanism-level separation axes appear in Mechanism of 4D-DIA Quantitative Proteomics.
When Label-Free 4D-DIA Fits Versus Other Quant Routes
Label-free 4D-DIA more often fits open discovery with staggered enrollment, many separate injections, and a relative matrix claim across groups.
Isobaric tandem mass tag (TMT) or isobaric tags for relative and absolute quantitation (iTRAQ) more often fit closed multi-arm sets labeled the same day with reporter readout inside a multiplex. That path is not the default label-free 4D-DIA workflow.
Parallel reaction monitoring (PRM) more often fits when peptide names are frozen and remeasurement is the claim. Open 4D-DIA remains discovery-oriented until list status changes.
Mobility-free label-free DIA or SWATH-style schedules may fit when pilots show clean extraction without mobility gains (SWATH Proteomics, SWATH-MS Label-Free Protein Quantification Technology Analysis).
Closed-list confirmation on mobility platforms compares with 4D-DIA vs 4D-PRM: Which 4D Quantitative Approach Suits Your Project Better.
End-to-End Label-Free 4D-DIA Workflow
A practical label-free 4D-DIA spine usually runs as follows.
- Confirm matrix, contrast, relative claim, and platform scope.
- Prepare matched digests with shared lysis, digestion, and cleanup (Wisniewski et al. 2009).
- Desalt to injection-ready loads without isobaric tagging.
- Pilot mobility-aware DIA; lock gradient, mobility plan, and window table.
- Acquire cohort files with interleaved run order when file count allows.
- Extract peptides under unified FDR and interference rules.
- Normalize, apply missing-value policy, roll up proteins, test predefined contrast only.
- Optional PRM or SIS on a short list after QC.
Steps 1 through 3 are label-free prep. Steps 4 through 6 are 4D-DIA acquisition and extraction. Step 7 is the quantitative claim layer.
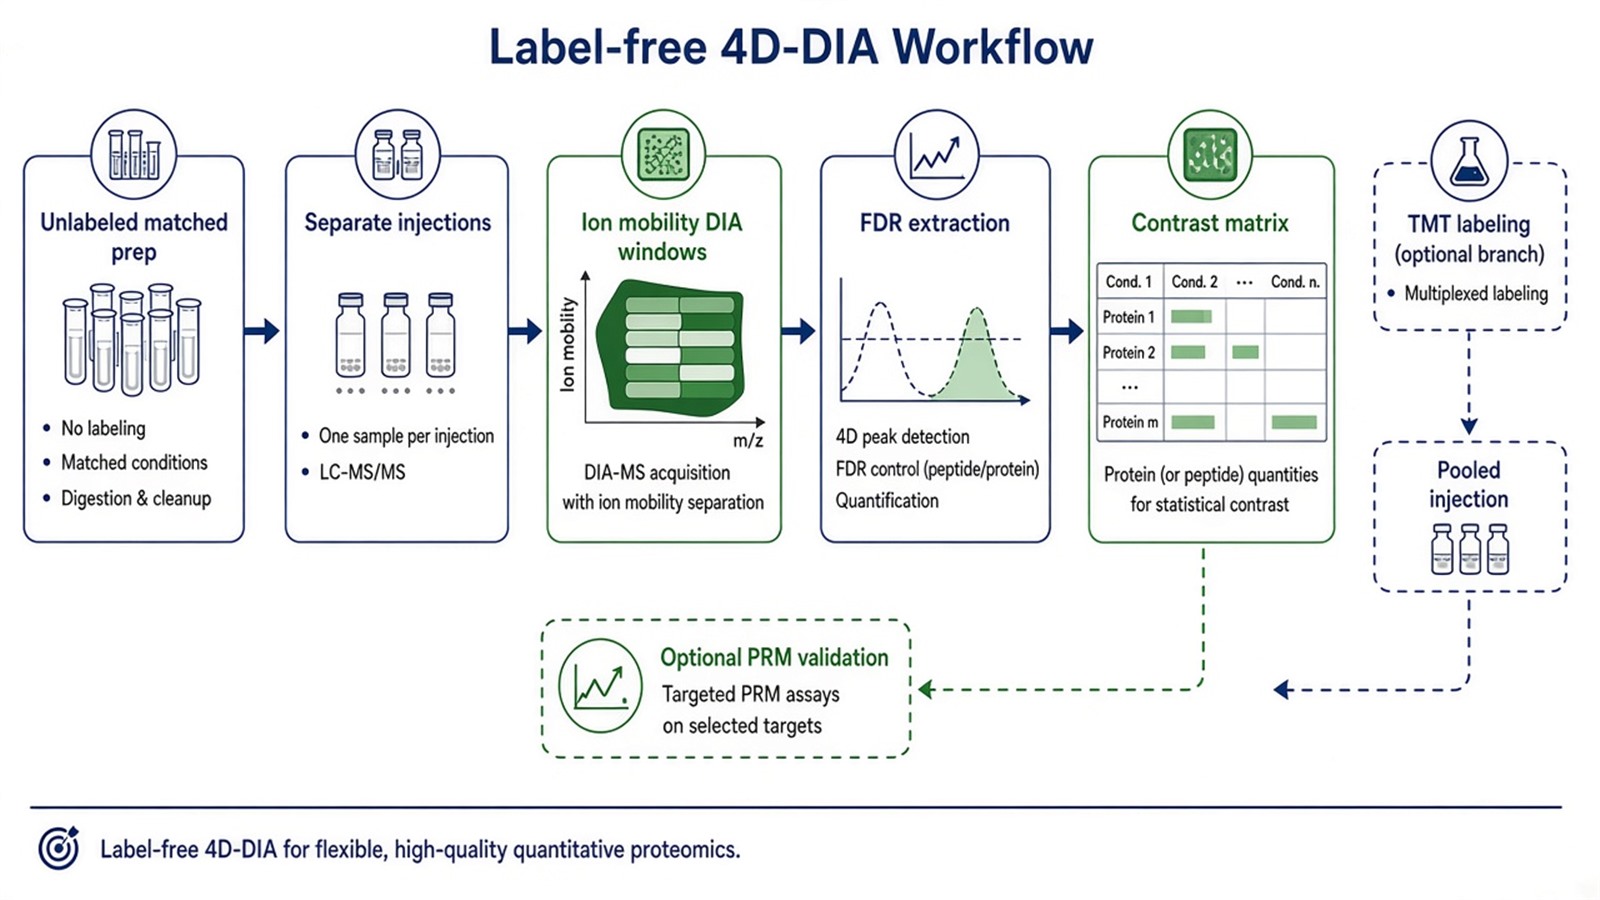
Figure 1. Label-free 4D-DIA couples unlabeled separate injections to mobility-enhanced DIA and declared extraction before contrast claims.
Operational detail appears in Workflow of 4D-DIA Quantitative Proteomics. Tradeoffs appear in Advantages and Disadvantages of 4D-DIA Quantitative Proteomics. Extraction conventions appear in DIA Data Analysis Methods in Proteomics.
Parameters and Limits in Label-Free 4D-DIA Projects
Topic |
Label-free 4D-DIA sensitivity |
Failure mode if ignored |
|---|---|---|
Chemistry |
Unlabeled separate injections |
Tagged and untagged files merged |
Prep |
Matched across arms |
Load bias mimics biology |
Mobility and windows |
Locked mid cohort |
Split incompatible matrices |
Extraction |
One declared version |
Mixed pipelines |
FDR |
Unified across files |
Drifting cutoffs |
Missing values |
Documented policy |
Imputed gaps without rules |
Claim |
Relative contrast default |
Open intensities as amount |
Limits follow the same table. Label-free 4D-DIA does not ensure every low-abundance protein in every matrix. Missing values often track prep loss and abundance, not only mobility width (Lazar et al. 2016). Forced 4D scheduling on simple matrices may add cost without pilot evidence.
Enrollment and pilot checks appear in Exploring Quantitative Analysis in Proteomics.
How to Choose Label-Free 4D-DIA for the Next Study
Use enrollment, pilot interference, and follow-up goal together.
Project need |
Label-free 4D-DIA plan that often fits |
Recheck before committing |
|---|---|---|
Open cohort, staggered samples |
Unlabeled 4D-DIA separate injections |
Same-day TMT assumption |
Complex digest, mobility pilot gain |
Lock 4D after pilot |
4D without interference evidence |
Simple matrix, clean mobility-free pilot |
Mobility-free label-free DIA |
Forced 4D complexity |
Closed same-day multiplex |
TMT or iTRAQ route |
Label-free 4D mixed with tags |
Short list after discovery |
Same prep, PRM panel |
Discovery p-values as final assay |
Amount on named peptides |
Targeted SIS assay |
Label-free 4D as concentration |
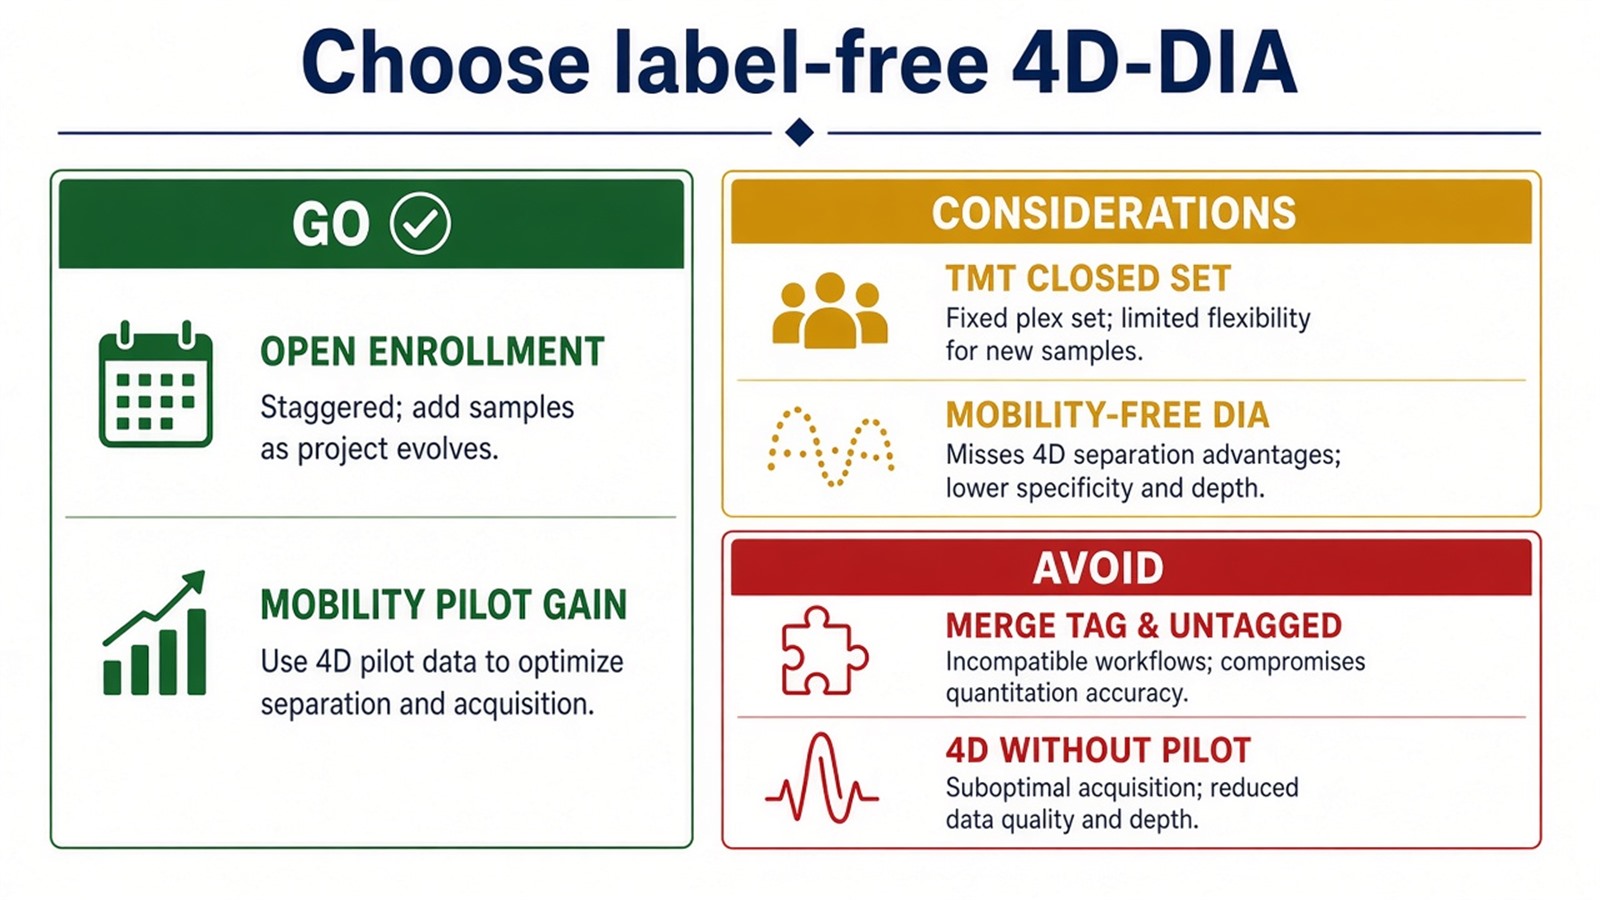
Figure 2. Choose label-free 4D-DIA when unlabeled separate injections and mobility-aware pilots fit; TMT, mobility-free DIA, or PRM fit other list and enrollment patterns.
Three-axis selection appears in Protein Quantification Method.
Teams that already have matrix, contrast, enrollment plan, and pilot scope can review the MtoZ Biolabs Label-Free Quantitative Proteomics Service for label-free 4D-DIA planning.
Frequently Asked Questions
1. Is label-free 4D-DIA the same as SWATH?
SWATH is a windowed DIA schedule. Label-free names the chemistry. A project can be label-free SWATH on mobility-free platforms or label-free 4D-DIA when mobility is used.
2. Does label-free 4D-DIA require TMT?
No for standard discovery on unlabeled injections. TMT is a separate isobaric route.
3. Can labeled and unlabeled files be analyzed as one contrast?
No. Chemistry and readout differ. Keep contrasts within one quantitative route.
4. Does 4D-DIA label-free data support diagnostic claims alone?
No. Discovery matrices need validation designs outside open label-free contrasts.
5. When should mobility-free DIA be kept?
When pilots show adequate extraction without mobility scheduling gains and operations favor simpler lock.
MtoZ Biolabs can support label-free 4D-DIA planning when sample type, contrast, enrollment flexibility, and intended relative claim are documented.
If further evaluation of prep matching, mobility pilots, extractor choice, or discovery-to-targeted staging is needed, project details can be submitted for a feasibility review.
Reference
- S.A. Gerber, J. Rush, O. Stemman, M.W. Kirschner, S.P. Gygi (2003). Absolute quantification of proteins and phosphoproteins from cell lysates by tandem MS. Proc. Natl. Acad. Sci. U.S.A., 100, 6940-6945. https://doi.org/10.1073/pnas.0832254100
- L.C. Gillet, P. Navarro, S. Tate, H. Rost, N. Selevsek, L. Reiter, R. Bonner, R. Aebersold (2012). Targeted data extraction of the MS/MS spectra generated by data-independent acquisition: a new concept for consistent and accurate proteome analysis. Mol. Cell. Proteomics, 11, O111.016717. https://doi.org/10.1074/mcp.O111.016717
- F. Meier, A.D. Brunner, O. Frank, M.T. Krauss, M. Koch, H. Teichert, C. Trede, J. Kosinski, T. Park, M. The, M. Mann (2020). diaPASEF: parallel accumulation-serial fragmentation combined with data-independent acquisition. Nat. Methods, 17, 1229-1236. https://doi.org/10.1038/s41592-020-00998-0
- V. Demichev, C. Messner, S. Vernardis, K.S. Lilley, M. Ralser (2020). DIA-NN: neural networks and interference correction enable deep proteome coverage in high throughput. Nat. Methods, 17, 41-44. https://doi.org/10.1038/s41592-019-0638-x
- C. Lazar, L. Gatto, M. Ferro, C. Bruley, T. Burger (2016). Accounting for the multiple natures of missing values in label-free quantitative proteomics data sets to compare imputation strategies. J. Proteome Res., 15, 1116-1125. https://doi.org/10.1021/acs.jproteome.5b00981
- G. Rosenberger, S. Liu, J. Röst, H.L. Ludwig, C. Buil, B. Borchers, R. Aebersold, L. Collins (2017). Statistical control of peptide and protein error rates in large-scale targeted data-independent acquisition analyses. Nat. Commun., 8, 14224. https://doi.org/10.1038/ncomms14224
- J.R. Wisniewski, A. Zougman, N. Nagaraj, M. Mann (2009). Universal sample preparation method for proteome analysis. Nat. Methods, 6, 359-362. https://doi.org/10.1038/nmeth.1322
