



Exploring quantitative analysis in proteomics starts with sample decisions that keep later peptide signals comparable across conditions. Matrix choice, storage, aliquot planning, and matched pretreatment decide whether an exploratory label-free, isobaric, or targeted run can support a relative claim, or a stable isotope-labeled internal standard (SIS) amount (Bantscheff et al. 2012). The guide below focuses on sample readiness and pretreatment for first-pass quantitative exploration. It is not a disease-application catalog, not a method-definition encyclopedia, and not a fixed price guide.
Exploration still has to name labeling chemistry and acquisition mode before digestion freezes. Unlabeled digests can be acquired as data-dependent acquisition (DDA) or data-independent acquisition (DIA). Isobaric tandem mass tag (TMT) work needs amine-compatible peptides and a freezable plex. Named proteins followed by parallel reaction monitoring (PRM) or multiple reaction monitoring (MRM) need clean digests and a closed peptide list. In MtoZ Biolabs offerings, unlabeled discovery is commonly run as DDA label-free or DIA, multiplexed isobaric work is typically evaluated as TMT, and closed lists are typically followed by PRM, with MRM when a scheduled triple-quadrupole panel already exists.
Sample Decisions That Shape Quantitative Analysis Up Front
Before lysis chemistry is chosen, four sample decisions usually decide whether exploration will be interpretable.
Enrollment flexibility. Open cohorts that may still add samples more often fit unlabeled separate injections. Sets that can be frozen now more often fit TMT mixing after digestion.
Matrix family. Cells, tissues, plasma or serum, and purified fractions need different disruption, depletion, and input rules. Mixing matrices inside one exploratory contrast without a declared adaptation usually breaks comparability.
Storage and freeze-thaw. Matched freeze history across groups matters as much as matched wet weight. Unequal thaw cycles create apparent abundance shifts before LC-MS starts.
Aliquot and backup plan. Exploration often needs a pilot digest plus a reserved aliquot for a second prep if cleanup fails. Consuming the only vial on an untested recipe raises project risk.
Sample readiness for quantitative exploration is a matched enrollment, matrix, storage, and aliquot plan, not a longer method keyword list.
Bulk Bradford, bicinchoninic acid (BCA), Lowry, and 280 nm assays help match total protein input. They do not replace peptide-level quantification after digestion.
Pretreatment Checks Before the First Exploration Run
First-pass exploration should lock a short pretreatment checklist before large batches.
Matched input. Aliquot by wet weight, cell count, or total protein under one load assay. Group-specific under-loading looks like biology later.
One lysis family per matrix. Share denaturation, reduction, and alkylation timing across the exploratory set. Filter-aided or detergent-compatible cleanup helps remove chaotropes and detergents that suppress LC-MS before trypsin digestion (Wisniewski et al. 2009).
Comparable cleanup and digestion. Peptide recovery after StageTip-style or solid-phase cleanup must stay matched (Rappsilber et al. 2007). Incomplete digestion that tracks treatment is not a discovery finding.
Route-specific checks. TMT needs amine-free peptide buffers before labeling. SIS amount claims need a declared spike point relative to digestion and cleanup (Gerber et al. 2003). Targeted exploration still needs unique peptides that fit one duty cycle.
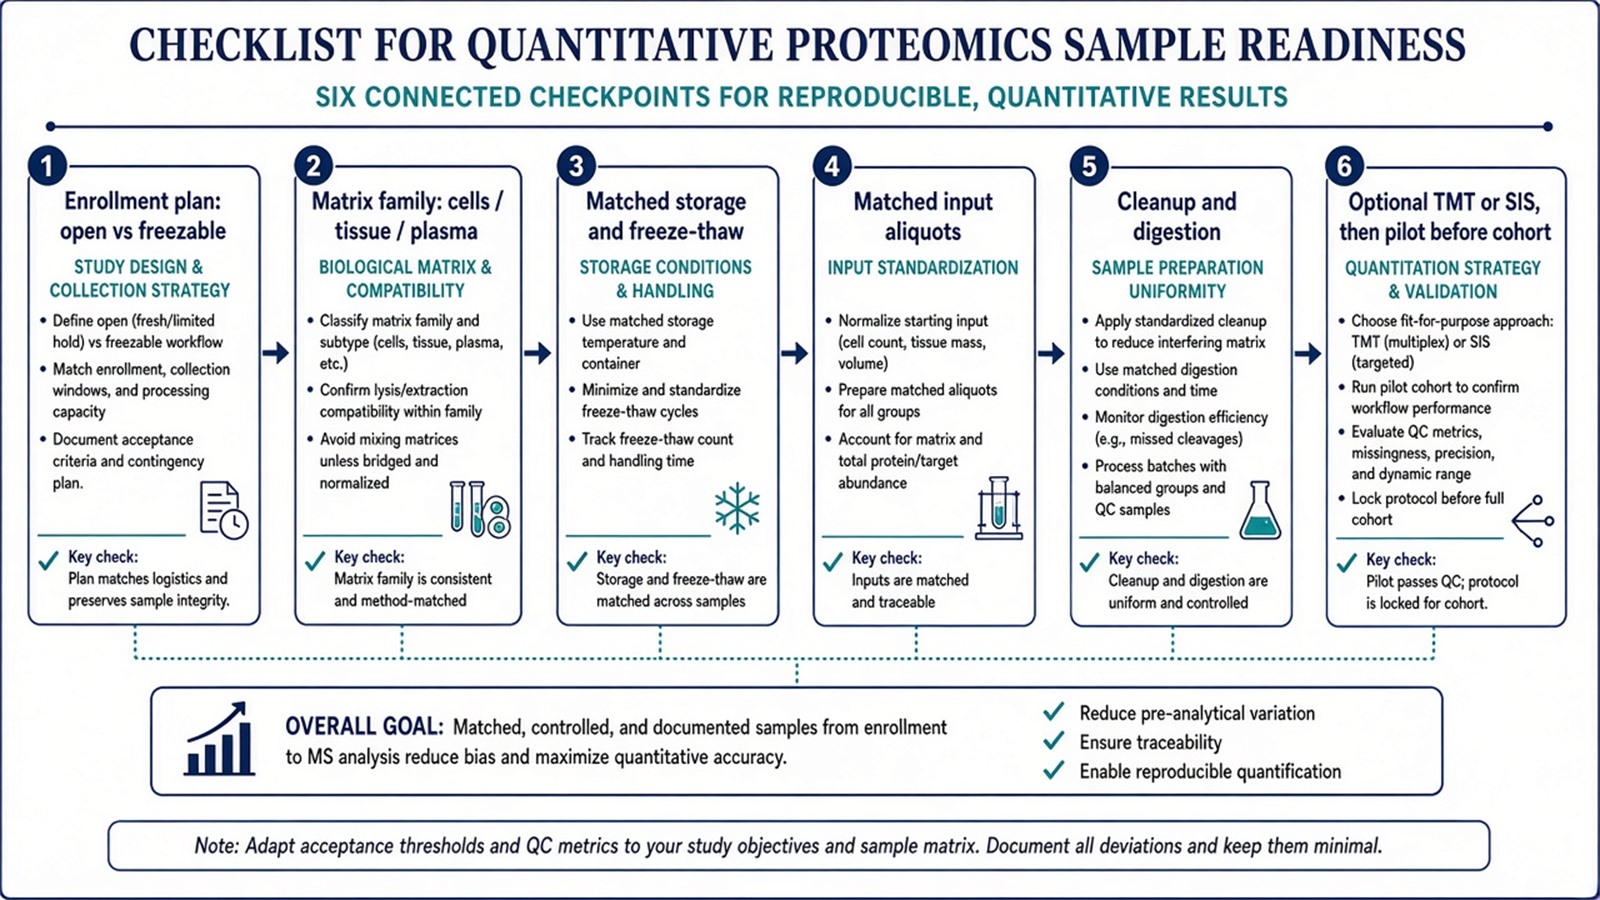
Figure 1. Exploratory quantitative analysis depends on matched sample readiness and pretreatment before the first LC-MS batch.
Researchers who need the full lysis-to-digest chemistry spine after this checklist can refer to Quantitative Protein Analysis.
Shared Sample-to-Digest Workflow for Exploratory Cohorts
A practical exploration spine usually runs as follows.
- Confirm matrix, enrollment plan, group map, and relative versus SIS amount claim.
- Reserve pilot and backup aliquots; record freeze-thaw history.
- Match input, then lyse or extract under one declared chemistry.
- Reduce, alkylate, digest, and clean under shared rules.
- Add TMT or SIS only when the exploration design requires them.
- Desalt a pilot set first; review recovery before the remaining cohort.
Where samples diverge still matters. Unlabeled exploration keeps each digest on its own injection after pretreatment. TMT mixes labeled digests after labeling. Targeted exploration freezes names first, then schedules peptides. Running the full exploratory cohort before a pilot cleanup check often forces a second digestion batch.
Order errors erase exploration value early. Unequal storage, treatment-confounded lysis batches, amine carryover before TMT, or SIS spiked into a group-biased digest can break the claim before acquisition.
Missing values later in the matrix often begin as pretreatment dropout, not only as acquisition chance (Lazar et al. 2016).
Parameters and Limits That Bound Exploration Prep
Parameter |
Why it matters in exploration |
Failure mode if ignored |
|---|---|---|
Matched input and freeze history |
Keeps peptide yield comparable |
Apparent fold changes from load or storage bias |
Single matrix family per contrast |
Stabilizes lysis and cleanup rules |
Mixed-matrix artifacts read as biology |
Detergent or chaotrope removal |
Protects chromatography and ionization |
Suppressed peaks in the pilot set |
Digestion completeness |
Stabilizes peptide stoichiometry |
Missed cleavages that track groups |
Amine-free buffer before TMT |
Allows channel labeling |
Soft reporters and incomplete plexes |
SIS spike timing |
Supports amount claims |
Relative peaks read as concentration |
Pilot-before-cohort rule |
Limits rework cost |
Full batch digested on an untested recipe |
Limits follow the same table. Exploration does not invent unidentified proteins. TMT cannot accept late unlabeled harvests as extra channels. Targeted exploration does not survey an open proteome. Ordinary discovery intensities after cleanup are still not ng per mL without SIS and calibration. Disease keywords without matched groups are not a sample plan.
Researchers who need signal-type definitions after sample readiness is fixed can refer to What Are the Quantitative Methods for Proteomics?.
Choosing a Sample Plan for the Next Exploration Project
Use enrollment flexibility, matrix, and claim unit to choose the first sample plan.
Project need |
Sample plan that often fits |
Recheck before committing |
|---|---|---|
Open enrollment, cells or tissue |
Matched unlabeled lysis and digest; separate injections |
Run order identical to treatment |
Full set freezable now |
Matched digest plus TMT labeling and mix |
Later unlabeled files; amine carryover |
Plasma or serum first look |
Equal depletion or fractionation plus matched digest |
Depletion on only one group |
Named peptides to remeasure |
Clean digest supporting a PRM or MRM panel |
Panel longer than one duty cycle |
Mass or molar unit required |
Matched digest plus assigned SIS and calibration |
Spike after incomplete digestion |
Only total load matching |
Bulk dye or A280 assay |
Not peptide-level exploration |
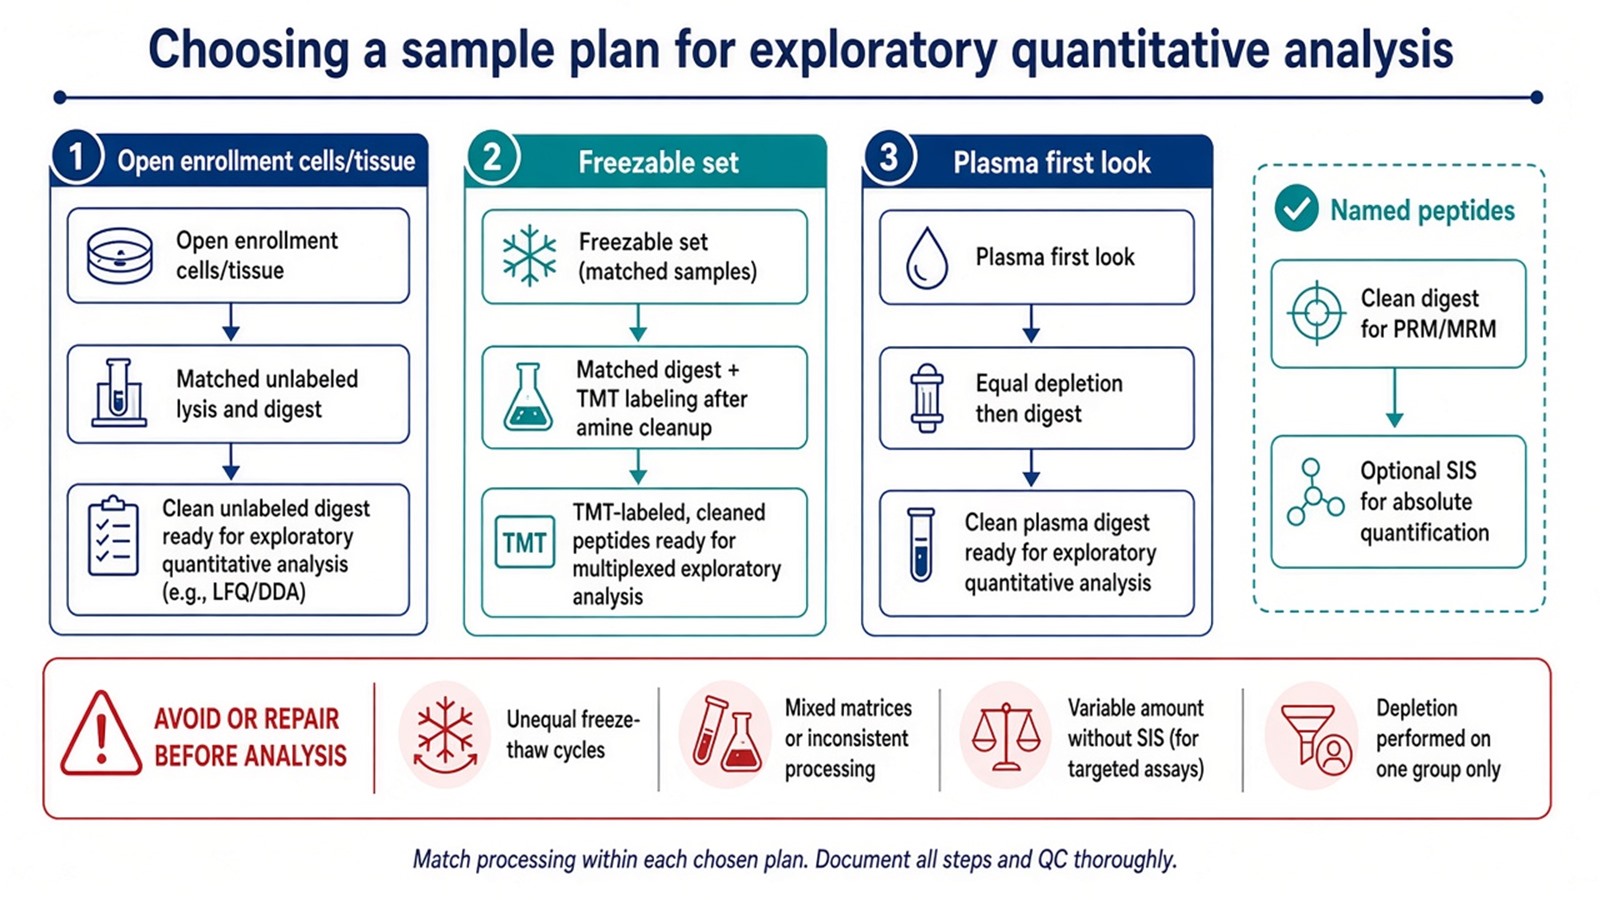
Figure 2. Choose the exploration sample plan from enrollment and claim unit; unequal storage, mixed matrices, and amount-without-SIS requests need repair first.
Unlabeled sample plans more often fit open exploration. TMT plans more often fit freezable multi-group sets after amine cleanup. Targeted confirmation more often fits after names freeze. Exceptions exist when a tiny validated panel never needed discovery, or when a short closed cohort must share one injection to limit run-order drift.
Researchers who need labeling-versus-acquisition selection after the sample plan is clear can refer to Protein Quantification Method. Teams that already have matrix, enrollment plan, and a requested claim can also review the MtoZ Biolabs Quantitative Proteomics Service for project-specific sample-readiness planning.
Frequently Asked Questions
1. Can exploration start from a bulk protein assay alone?
No. Bulk assays help match input. Peptide-level quantitative exploration still needs matched pretreatment and a declared claim.
2. Should every exploratory project deplete plasma?
No. Depletion can help deep discovery when applied equally across groups. Unequal depletion creates artificial contrasts and is not required for every first look.
3. How many freeze-thaw cycles are acceptable before exploration?
Keep freeze-thaw history matched across groups and minimize cycles. Unequal thaw counts are a design problem even when total protein looks matched.
4. Is a pilot digest required before the full exploratory cohort?
Usually yes when the recipe or matrix is new to the lab. Digestion of the entire set on an untested cleanup path often forces costly repeats.
5. Do unlabeled and TMT exploration share the same sample plan?
They share matched input, lysis, digestion, and cleanup principles. TMT adds amine-compatible labeling and mixing after digestion. Unlabeled plans keep digests on separate injections.
MtoZ Biolabs can support exploratory quantitative sample planning across unlabeled DDA or DIA discovery, TMT relative plexes, and targeted PRM or MRM follow-up when sample type, study design, and the intended claim are available.
If further evaluation of enrollment flexibility, matrix-matched lysis, depletion needs, or SIS spike planning is needed, project details can be submitted for a feasibility review.
Reference
- M. Bantscheff, S. Lemeer, M.M. Savitski, B. Kuster (2012). Quantitative mass spectrometry in proteomics: critical review update from 2007 to the present. Anal. Bioanal. Chem., 404, 939-965. https://doi.org/10.1007/s00216-012-6203-4
- S.A. Gerber, J. Rush, O. Stemman, M.W. Kirschner, S.P. Gygi (2003). Absolute quantification of proteins and phosphoproteins from cell lysates by tandem MS. Proc. Natl. Acad. Sci. U.S.A., 100, 6940-6945. https://doi.org/10.1073/pnas.0832254100
- C. Lazar, L. Gatto, M. Ferro, C. Bruley, T. Burger (2016). Accounting for the multiple natures of missing values in label-free quantitative proteomics data sets to compare imputation strategies. J. Proteome Res., 15, 1116-1125. https://doi.org/10.1021/acs.jproteome.5b00981
- J. Rappsilber, M. Mann, Y. Ishihama (2007). Protocol for micro-purification, enrichment, pre-fractionation and storage of peptides for proteomics using StageTips. Nat. Protoc., 2, 1896-1906. https://doi.org/10.1038/nprot.2007.261
- J.R. Wisniewski, A. Zougman, N. Nagaraj, M. Mann (2009). Universal sample preparation method for proteome analysis. Nat. Methods, 6, 359-362. https://doi.org/10.1038/nmeth.1322
