



Four-dimensional data-independent acquisition (4D-DIA) quantitative proteomics combines liquid chromatography (LC) separation, ion mobility separation, windowed MS/MS, and intensity-based extraction into one project record (Meier et al. 2020). In common platform language, diaPASEF-style schedules accumulate precursors in the mobility dimension and fragment them in synchronized m/z windows. Quantification is not read directly from the mass spectrometer display. It comes from extracted fragment chromatograms under false discovery rate (FDR) control after digests are acquired on label-free separate injections in most discovery work (Gillet et al. 2012). The guide below explains those mechanisms in project terms and ties them to workflow, parameters, and fit. It is not a physics textbook, not a vendor tuning sheet, and not a prospective applications catalog.
Ion mobility spreads co-eluting precursors in collision cross section space before MS/MS. That can reduce overlapping fragment signatures in crowded digests when methods are locked. Mobility does not remove matched prep, frozen contrasts, or missing-value rules (Rosenberger et al. 2017; Lazar et al. 2016).
In MtoZ Biolabs offerings, mobility-enhanced DIA is discussed when platform scope supports ion mobility-aware acquisition and extraction on the study matrix.
What the Four Dimensions Mean in 4D-DIA Projects
Project records should name four separable information layers, not marketing labels.
LC retention time orders peptide elution from the column. It sets peak width and how many precursors occupy each acquisition cycle.
Precursor m/z is the mass-to-charge axis used to define DIA windows and precursor masses in extraction.
Ion mobility adds gas-phase separation of ions with different collision cross sections at a given m/z. It is the usual meaning of the fourth dimension in 4D-DIA, not a substitute for chromatography time.
Fragment intensity traces over acquisition cycles supply the quantitative signal after extraction, normalization, and contrast testing.
4D-DIA mechanism language should separate separation axes from the quantitative claim, which remains a predefined relative contrast unless SIS targeting is declared (Gerber et al. 2003).
Label-free versus isobaric chemistry is independent of mobility (Is Labeling Required for 4D-DIA Quantitative Proteomics). Operational steps appear in Workflow of 4D-DIA Quantitative Proteomics.
Ion Mobility and DIA Window Fragmentation
Mechanisms at acquisition time stack as follows.
Peptides elute from LC and are ionized. Ions enter a mobility cell where nitrogen or other buffer gas separates packets by mobility.
In diaPASEF-style plans, selected mobility slices release precursors for windowed MS/MS while other ions continue accumulating, which couples mobility duty cycle with DIA window scheduling (Meier et al. 2020).
Each DIA window fragments many precursors in that m/z range per cycle. The instrument stores MS/MS traces. It does not intensity-pick individual precursors in real time the way data-dependent acquisition (DDA) does.
Window width, cycle time, and mobility scheduling must fit elution density. Over-wide windows increase fragment overlap. Over-narrow windows leave m/z gaps.
Open discovery keeps the protein list unconstrained at acquisition. Closed lists move to parallel reaction monitoring (PRM) routes (4D-DIA vs 4D-PRM: Which 4D Quantitative Approach Suits Your Project Better).
How Quantification Emerges from Extracted Fragment Traces
Acquisition mechanisms stop at stored spectra. Quantification mechanisms continue in software.
Library-based extraction matches observed fragments to expected transitions from spectral libraries. Library-free search identifies peptides under FDR control without a separate library campaign (Demichev et al. 2020).
Mobility-aware extractors use ion mobility as an additional match constraint when the pipeline supports it. That can reject ambiguous peak groups when co-elution is high.
Peak groups integrate to intensities per run. Normalization aligns runs. Protein rollup applies a declared rule. Only the predefined contrast is tested statistically.
Missing values often track low abundance, interference, and prep loss, not only window design (Lazar et al. 2016).
General extraction principles appear in DIA Data Analysis Methods in Proteomics.
Mechanism Linked Workflow Spine
A practical sequence shows where mechanisms hand off.
- Confirm matrix, contrast, and relative claim.
- Prepare matched digests to injection-ready loads.
- Pilot mobility-aware DIA; lock LC, mobility, and window parameters.
- Acquire cohort under the locked plan.
- Extract under unified FDR and interference filters.
- Normalize, handle missing values, test predefined contrast only.
- Optional PRM on a short list when list status changes.
Steps 1 through 2 are prep mechanisms outside 4D-DIA naming. Steps 3 through 4 are separation and fragmentation mechanisms. Steps 5 through 6 are quantification mechanisms.
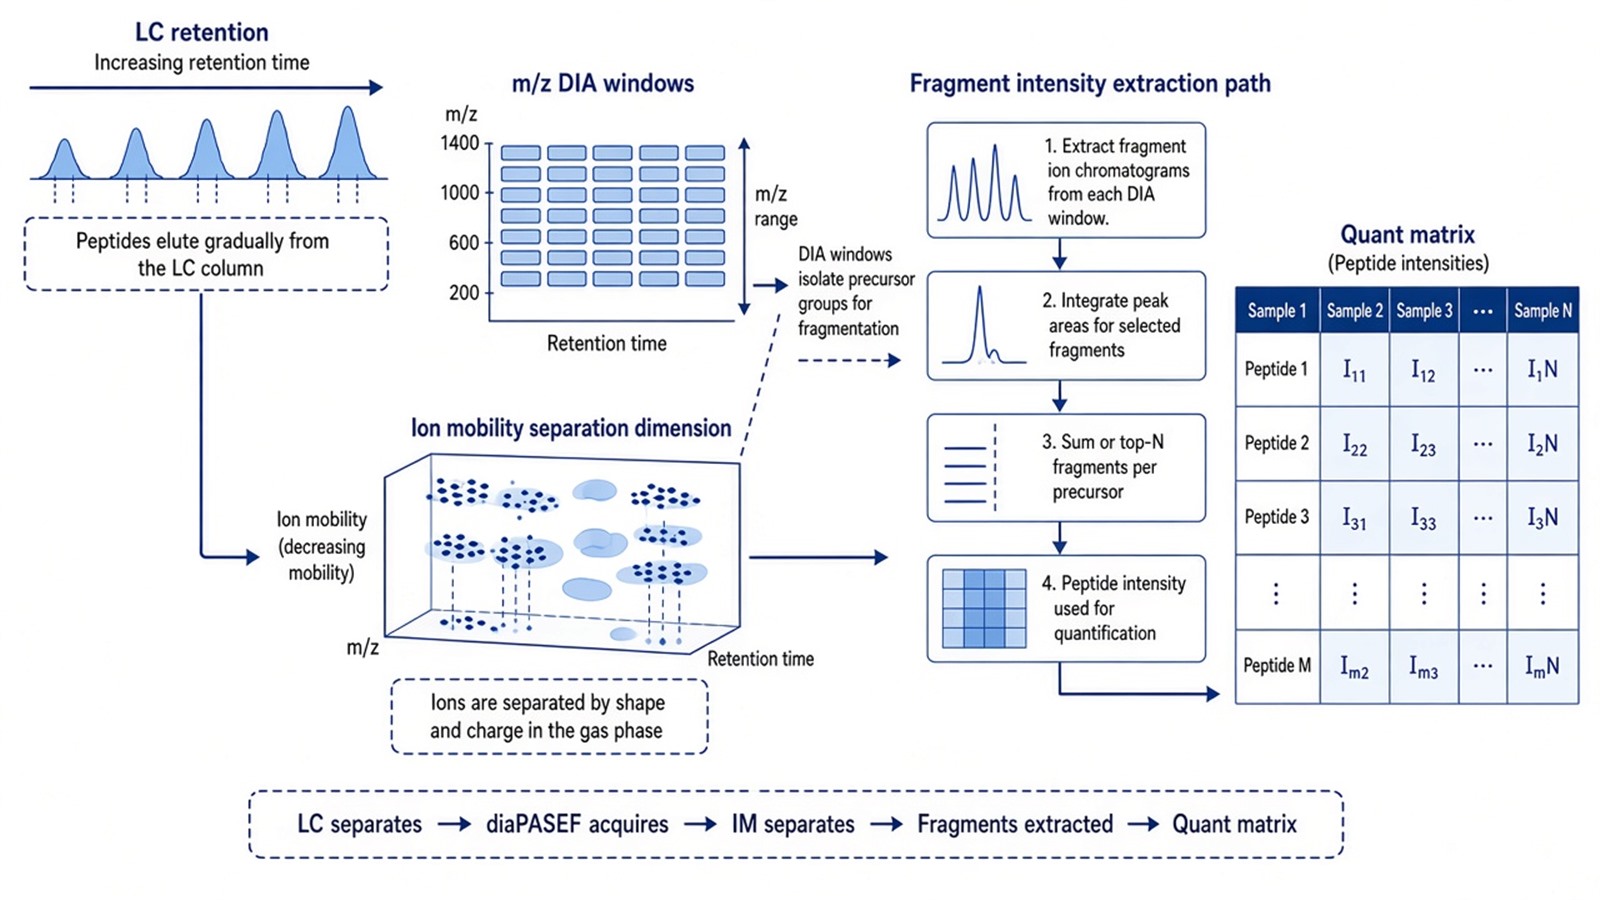
Figure 1. 4D-DIA couples mobility-aware acquisition to extraction-based quantification; separation axes are not the final quantitative claim.
Tradeoffs that affect whether mechanisms pay off appear in Advantages and Disadvantages of 4D-DIA Quantitative Proteomics.
Parameters and Limits Tied to 4D-DIA Mechanisms
Mechanism layer |
Parameter that must be declared |
Limit if mis-set |
|---|---|---|
LC |
Gradient length and load |
Cycle under-sampling |
Mobility |
Cell parameters and scheduling |
Invalid mobility match |
DIA windows |
Width, count, cycle time |
Overlap or coverage gaps |
Extraction |
Library provenance or library-free route |
Wrong search space |
FDR |
Unified model across files |
Drifting error control |
Contrast |
Frozen before threshold tuning |
Post hoc relabeling |
Claim |
Relative discovery default |
Open intensities as amount |
Limits follow the same table. 4D-DIA mechanisms do not prove diagnostic utility or drug mechanism by themselves. Mobility does not ensure detection of every low-abundance protein. Forced 4D scheduling on simple matrices may add complexity without interference benefit.
Mobility-free DIA or SWATH-style programs remain valid when pilots show adequate extraction (SWATH Proteomics).
How to Choose 4D-DIA Based on Mechanistic Fit
Use pilot interference, list status, and operations together.
Observation in pilots |
Mechanism interpretation |
Plan that often fits |
|---|---|---|
High co-elution, ambiguous MS/MS |
Mobility may help separation |
Lock 4D-DIA after pilot |
Clean extraction mobility-free |
Extra axis may add little |
Mobility-free DIA |
Open contrast, large cohort |
Window plus mobility lock needed |
Cohort 4D with interleaving |
Closed peptide list |
Targeted isolation fits |
4D-PRM or PRM |
Amount on named peptides |
SIS targeted quant |
Not open 4D intensities |
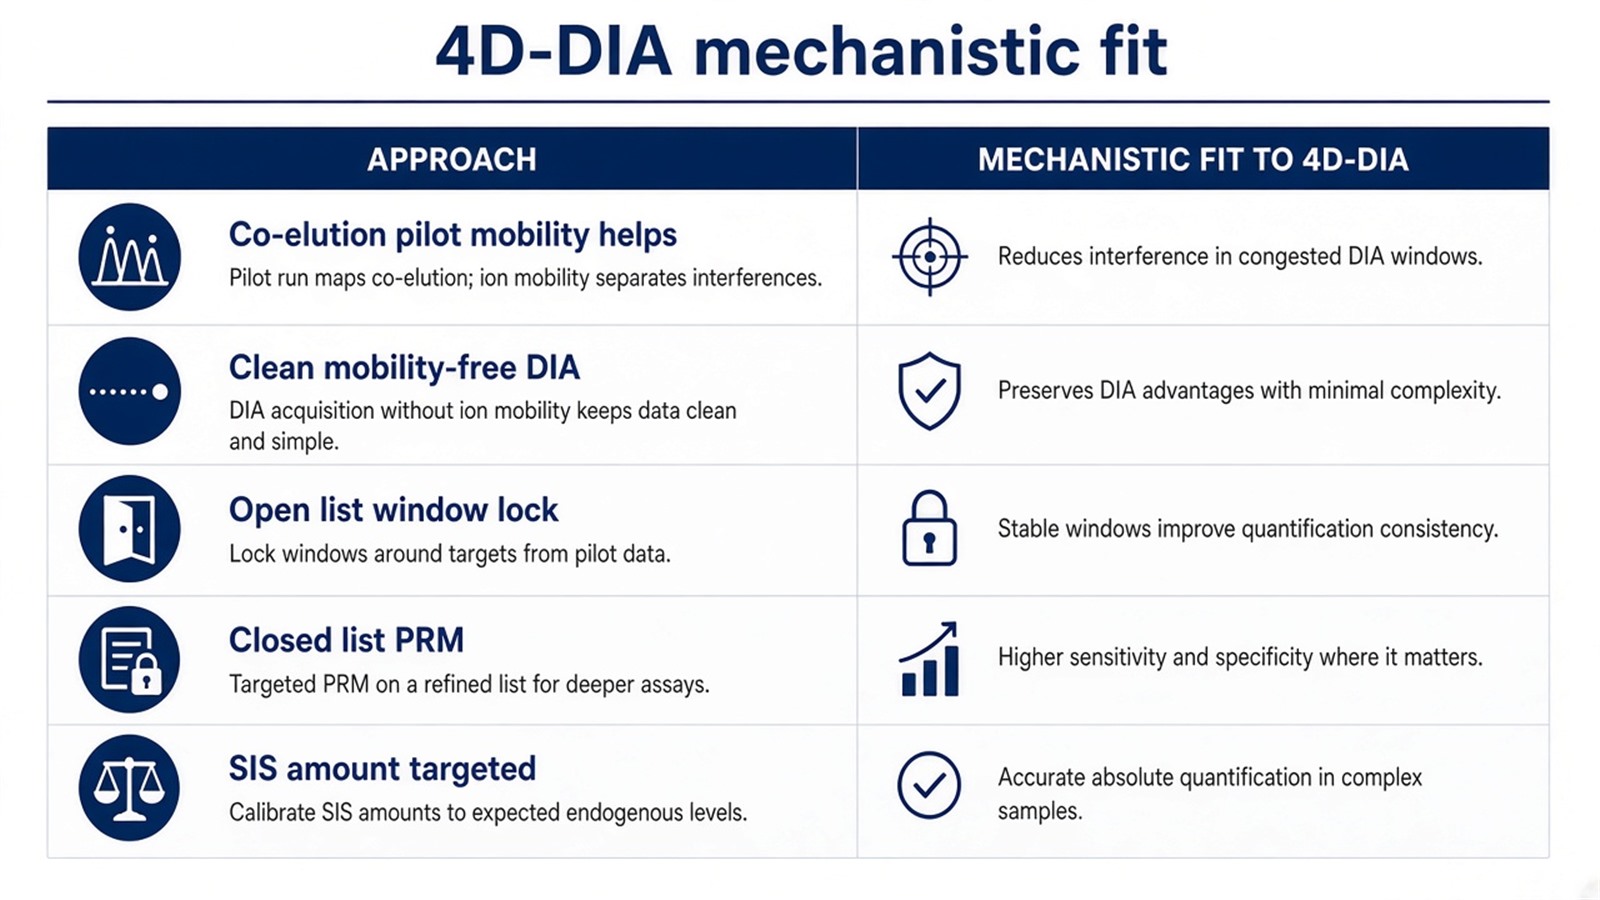
Figure 2. Choose 4D-DIA when mobility-aware pilots show separation gains on crowded MS/MS; otherwise mobility-free DIA or PRM may fit list status better.
Three-axis route selection appears in Protein Quantification Method. Staged confirmation appears in Application of DIA-PRM Proteomics.
Teams that already have pilot spectra, contrast design, and platform scope can review the MtoZ Biolabs Label-Free Quantitative Proteomics Service for 4D-DIA mechanism and method alignment.
Frequently Asked Questions
1. Does 4D-DIA mean the instrument autonomously picks peptides like DDA?
No. DIA fragments ions inside declared windows without intensity-based precursor picking during acquisition.
2. Is LC retention time the fourth dimension?
In 4D-DIA project language, the added dimension beyond conventional LC-MS/MS DIA is usually ion mobility, alongside retention time and m/z.
3. Does mobility replace FDR control?
No. FDR and interference filters remain mandatory in extraction.
4. Can mechanisms be changed mid cohort?
Changing mobility or window programs mid study splits comparability. New settings need a new block and analysis record.
5. Are more MS/MS scans the same as better quantification?
No. Identification sampling and contrast quality differ. Quantification needs stable extraction and replicate design.
MtoZ Biolabs can discuss 4D-DIA mechanisms in project planning when matrix pilots, contrast design, and platform capabilities are documented.
If further evaluation of mobility scheduling, window design, extractor choice, or discovery-to-targeted staging is needed, project details can be submitted for a feasibility review.
Reference
- S.A. Gerber, J. Rush, O. Stemman, M.W. Kirschner, S.P. Gygi (2003). Absolute quantification of proteins and phosphoproteins from cell lysates by tandem MS. Proc. Natl. Acad. Sci. U.S.A., 100, 6940-6945. https://doi.org/10.1073/pnas.0832254100
- L.C. Gillet, P. Navarro, S. Tate, H. Rost, N. Selevsek, L. Reiter, R. Bonner, R. Aebersold (2012). Targeted data extraction of the MS/MS spectra generated by data-independent acquisition: a new concept for consistent and accurate proteome analysis. Mol. Cell. Proteomics, 11, O111.016717. https://doi.org/10.1074/mcp.O111.016717
- F. Meier, A.D. Brunner, O. Frank, M.T. Krauss, M. Koch, H. Teichert, C. Trede, J. Kosinski, T. Park, M. The, M. Mann (2020). diaPASEF: parallel accumulation-serial fragmentation combined with data-independent acquisition. Nat. Methods, 17, 1229-1236. https://doi.org/10.1038/s41592-020-00998-0
- V. Demichev, C. Messner, S. Vernardis, K.S. Lilley, M. Ralser (2020). DIA-NN: neural networks and interference correction enable deep proteome coverage in high throughput. Nat. Methods, 17, 41-44. https://doi.org/10.1038/s41592-019-0638-x
- C. Lazar, L. Gatto, M. Ferro, C. Bruley, T. Burger (2016). Accounting for the multiple natures of missing values in label-free quantitative proteomics data sets to compare imputation strategies. J. Proteome Res., 15, 1116-1125. https://doi.org/10.1021/acs.jproteome.5b00981
- G. Rosenberger, S. Liu, J. Röst, H.L. Ludwig, C. Buil, B. Borchers, R. Aebersold, L. Collins (2017). Statistical control of peptide and protein error rates in large-scale targeted data-independent acquisition analyses. Nat. Commun., 8, 14224. https://doi.org/10.1038/ncomms14224
