Advantages and Disadvantages of 4D-DIA Quantitative Proteomics
Four-dimensional data-independent acquisition (4D-DIA) quantitative proteomics adds ion mobility separation before windowed MS/MS on liquid chromatography-tandem mass spectrometry (LC-MS/MS), so precursors are resolved by retention time, m/z, and mobility before fragmentation (Meier et al. 2020). Quantification still uses extracted fragment traces under false discovery rate (FDR) control on label-free digests, not mobility alone (Gillet et al. 2012). The guide below weighs practical advantages and disadvantages of 4D-DIA for project planning, then links tradeoffs to workflow, parameters, and fit. It is not a platform advertisement, not a DDA versus DIA tutorial, and not a substitute for a signed methods appendix.
Mobility can reduce co-elution interference in complex digests. It does not remove matched prep requirements, frozen contrasts, or missing-value policy after extraction (Rosenberger et al. 2017; Lazar et al. 2016).
In MtoZ Biolabs offerings, mobility-enhanced DIA discovery is evaluated when the platform and matrix support ion mobility-aware acquisition and extraction, alongside mobility-free DIA when cohort constraints favor simpler method lock.
What 4D-DIA Means Before Weighing Tradeoffs
4D-DIA keeps an open protein list with windowed DIA acquisition plus mobility filtering during extraction or acquisition scheduling, depending on platform workflow.
Label-free separate injections remain the common chemistry. Isobaric tagging is a different route; mobility does not force or forbid tags (Is Labeling Required for 4D-DIA Quantitative Proteomics).
Relative contrast across predefined groups is the default claim. Amount on named peptides needs stable isotope-labeled internal standard (SIS) targeted designs, not open 4D-DIA intensities alone (Gerber et al. 2003).
Tradeoffs below assume a declared contrast, matched prep, and one locked extractor version across the cohort.
Open discovery versus closed remeasurement compares with 4D-DIA vs 4D-PRM: Which 4D Quantitative Approach Suits Your Project Better.
Advantages That Matter in Real Cohorts
Several 4D-DIA strengths appear repeatedly in complex matrices when methods are locked early.
Extra separation before MS/MS can narrow candidate interference when many precursors co-elute in mobility-free DIA on dense digests (Meier et al. 2020).
Reproducible windowed acquisition still supports multi-file cohorts when run order and prep are matched, similar to mobility-free DIA programs.
Stored raw data can be re-extracted with a declared pipeline and contrast, though reprocessing does not fix unequal prep or relabeled groups.
Library-free or library-based extractors with mobility-aware scoring can improve confidence on crowded MS/MS when interference filters are documented (Demichev et al. 2020).
Mobility-aware 4D-DIA does not automatically increase protein count in every matrix. Gains are contrast- and matrix-dependent and need pilot QC.
Disadvantages and Operational Costs
4D-DIA carries costs that matter in feasibility review.
Method complexity rises. Window plans, cycle time, mobility settings, and extractor compatibility must be declared and kept stable mid cohort.
Compute and storage load often exceed mobility-free DIA when file size and search space grow. Analysis timelines need explicit planning.
Platform and expertise dependency is higher. Cross-lab comparability needs shared acquisition tables and extraction versions, not instrument brand names alone.
Low-abundance peptides can still drop out when load, ion suppression, or library mismatch limit traces. Mobility is not a substitute for input mass or depletion design.
Missing values and batch effects still require documented policy. 4D-DIA does not remove biological replicate requirements or validation cohorts for deployment claims.
Shared Workflow Where Pros and Cons Appear
A practical 4D-DIA spine shows where tradeoffs land.
- Confirm matrix, contrast, relative claim, and mobility platform scope.
- Prepare matched digests with shared cleanup rules.
- Pilot mobility-aware DIA on a subset; lock window, cycle, and mobility plan.
- Acquire cohort with interleaved run order when file count allows.
- Extract under unified FDR and interference rules with one extractor version.
- Normalize, handle missing values, test predefined contrast only.
- Optional parallel reaction monitoring (PRM) on a short list when confirmation is in scope.
Steps 2 and 3 decide whether 4D-DIA complexity pays off versus mobility-free DIA on the same matrix. Steps 5 and 6 are where compute cost and FDR discipline meet.
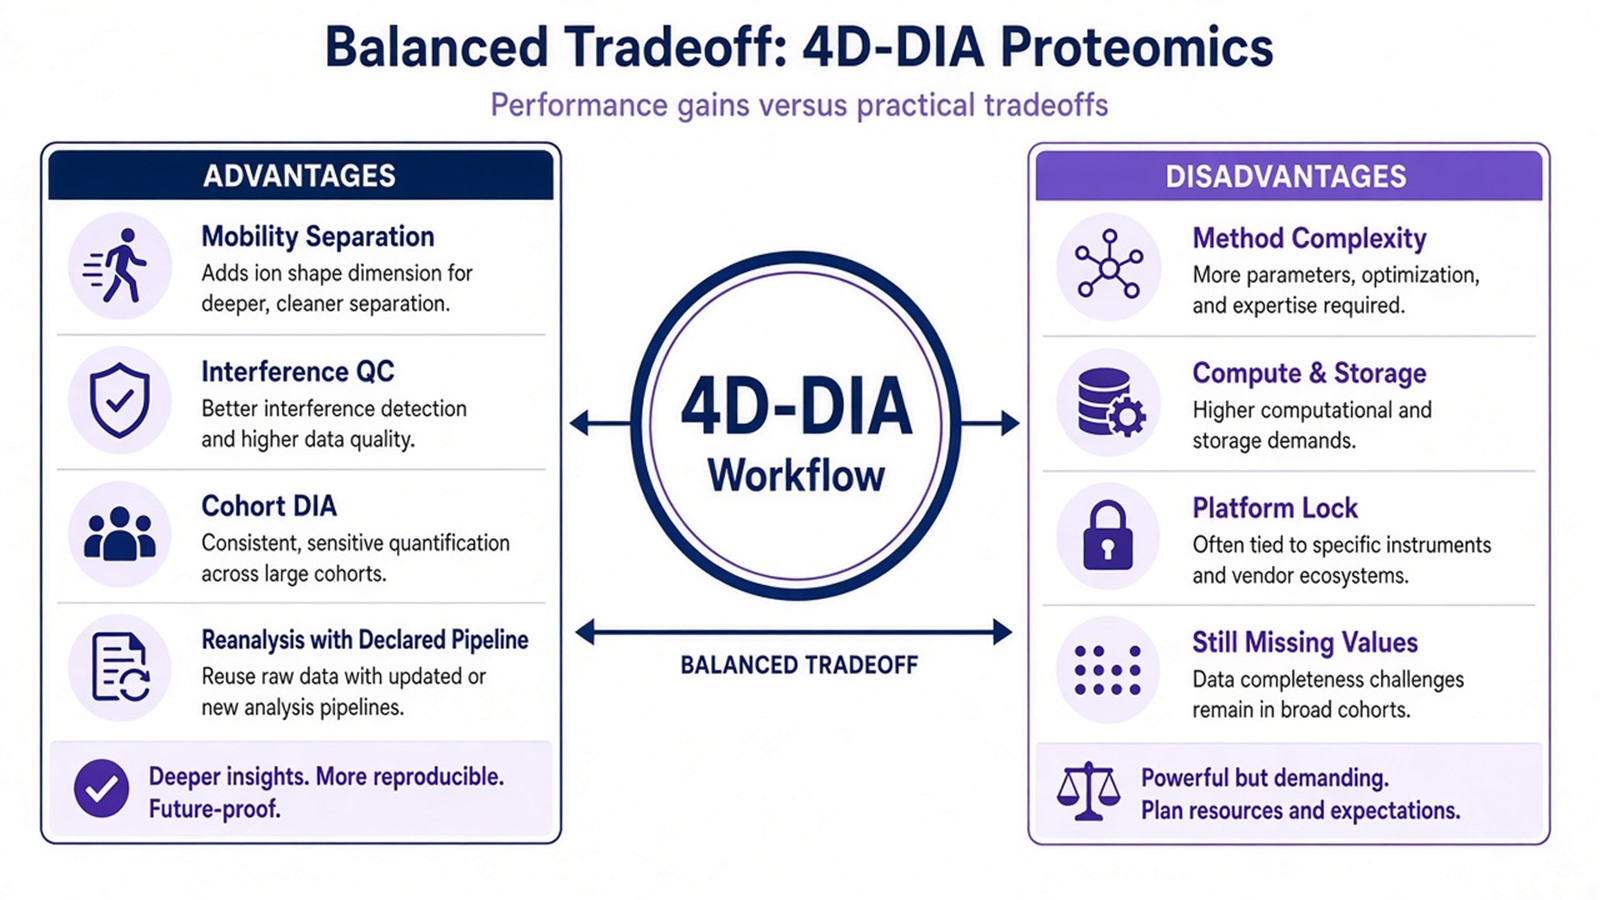
Figure 1. 4D-DIA tradeoffs cluster around method lock and extraction QC, not around mobility as a standalone benefit.
Extraction conventions appear in DIA Data Analysis Methods in Proteomics. General DIA and SWATH program context appears in SWATH Proteomics.
Parameters That Shift the Balance
Factor |
Tends to favor 4D-DIA |
Tends to favor mobility-free DIA or PRM |
|---|---|---|
Matrix complexity |
Dense co-elution in pilots |
Simple digests with clean pilots |
List status |
Open discovery contrast |
Closed list already defined |
Cohort size |
Many files, one locked 4D plan |
Small pilot or tight compute budget |
Multi-site need |
Shared method transfer documented |
Single-site with simpler lock |
Follow-up |
Discovery then PRM on short list |
PRM-only on known panel |
Claim unit |
Relative matrix |
SIS amount on named peptides |
Limits follow the same table. 4D-DIA does not certify diagnostics or drug labels. Forced 4D-DIA on abundant cell lines with easy prep may add cost without clear interference gain. PRM fits when names are frozen regardless of mobility.
How to Choose 4D-DIA Given Pros and Cons
Use matrix pilots, list status, and operational capacity together.
Project need |
Route that often fits |
Recheck before committing |
|---|---|---|
Complex digest, interference in mobility-free pilot |
4D-DIA with locked plan |
4D label without pilot evidence |
Large open contrast, platform supports 4D |
4D-DIA cohort after lock |
Mid-cohort mobility setting changes |
Tight compute or timeline |
Mobility-free DIA |
Full 4D reprocess cost unbudgeted |
Closed peptide panel |
4D-PRM or PRM |
Full 4D-DIA when list fixed |
Multi-site study |
4D-DIA only with transfer SOP |
Different extractors per site |
Amount on selected peptides |
Targeted SIS assay |
Open 4D-DIA as concentration |
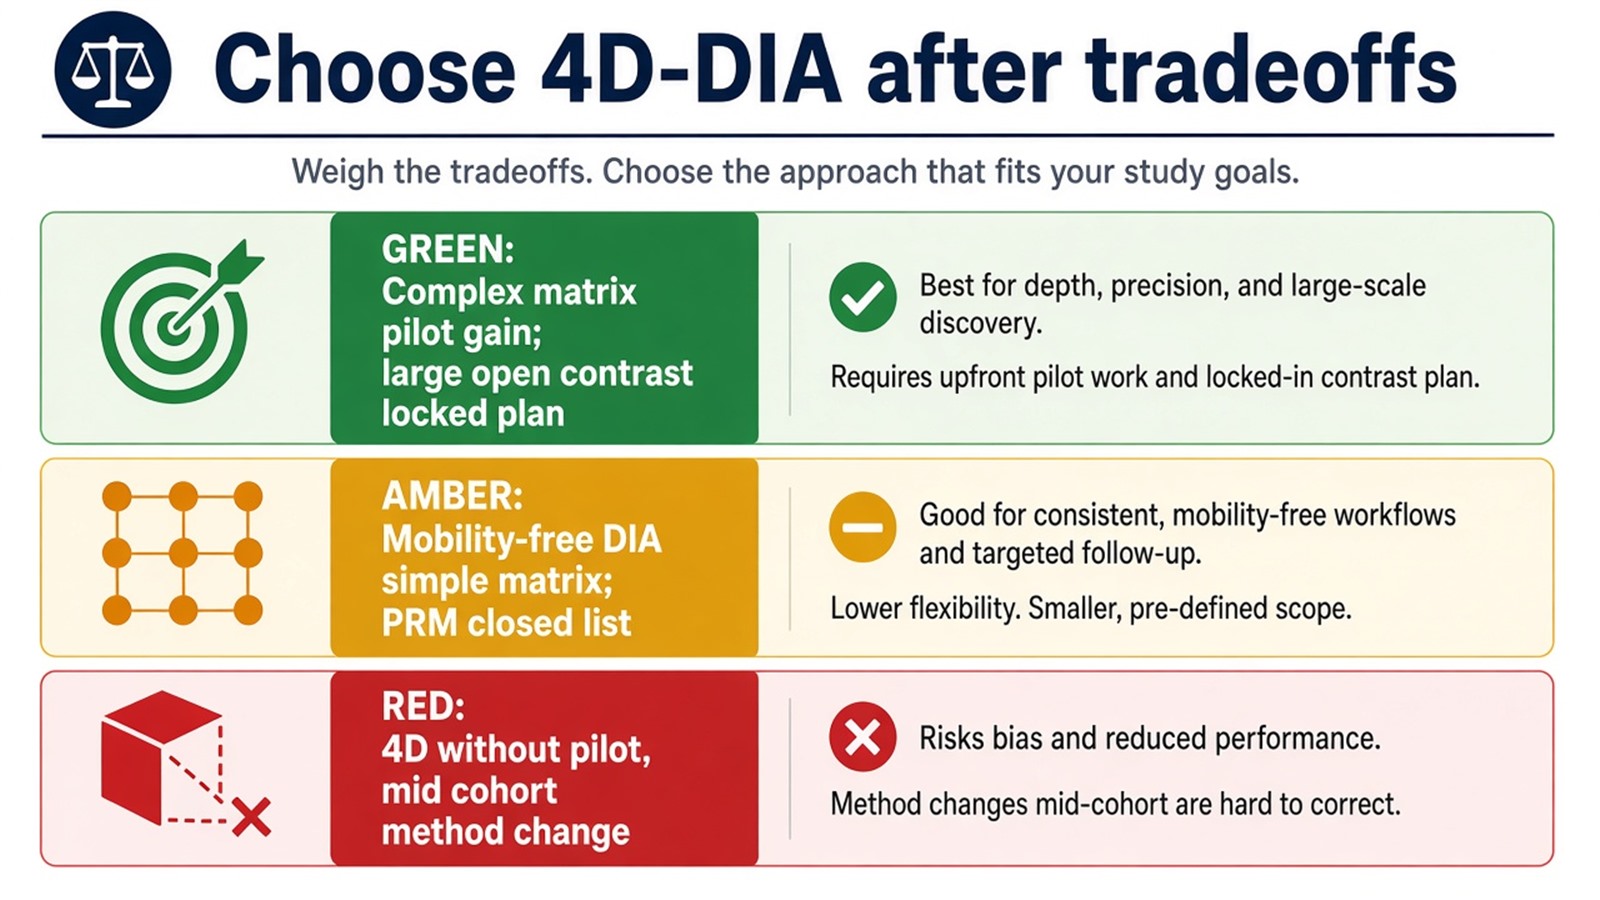
Figure 2. Choose 4D-DIA when mobility-aware pilots show interference gains; choose mobility-free DIA or PRM when list status or operations favor simpler lock.
Three-axis chemistry and acquisition selection appears in Protein Quantification Method. Staged confirmation appears in Application of DIA-PRM Proteomics.
Teams that already have matrix pilots, contrast, and operational constraints can review the MtoZ Biolabs Label-Free Quantitative Proteomics Service for 4D-DIA feasibility when the platform is in scope.
Frequently Asked Questions
1. Is 4D-DIA always better than mobility-free DIA?
No. Benefit depends on matrix interference and locked methods. Simple matrices may not justify added complexity.
2. Does 4D-DIA remove the need for PRM?
No. Open 4D-DIA remains discovery-oriented. Closed remeasurement still uses targeted designs when required.
3. Does ion mobility reduce sample amount requirements?
Not automatically. Input and prep still set peptide load into LC-MS.
4. Can 4D-DIA raw data be re-analyzed freely?
Re-extraction is possible with a declared pipeline and contrast. It does not fix prep or relabeling errors.
5. Are higher protein IDs the same as better quantitative decisions?
No. Identification depth and contrast quality differ. Missing-value rules and replicate design still decide strength.
MtoZ Biolabs can discuss 4D-DIA tradeoffs when matrix pilot data, contrast design, and platform scope are available.
If further evaluation of mobility pilots, window locking, extractor choice, or discovery-to-targeted staging is needed, project details can be submitted for a feasibility review.
Reference
- S.A. Gerber, J. Rush, O. Stemman, M.W. Kirschner, S.P. Gygi (2003). Absolute quantification of proteins and phosphoproteins from cell lysates by tandem MS. Proc. Natl. Acad. Sci. U.S.A., 100, 6940-6945. https://doi.org/10.1073/pnas.0832254100
- L.C. Gillet, P. Navarro, S. Tate, H. Rost, N. Selevsek, L. Reiter, R. Bonner, R. Aebersold (2012). Targeted data extraction of the MS/MS spectra generated by data-independent acquisition: a new concept for consistent and accurate proteome analysis. Mol. Cell. Proteomics, 11, O111.016717. https://doi.org/10.1074/mcp.O111.016717
- F. Meier, A.D. Brunner, O. Frank, M.T. Krauss, M. Koch, H. Teichert, C. Trede, J. Kosinski, T. Park, M. The, M. Mann (2020). diaPASEF: parallel accumulation-serial fragmentation combined with data-independent acquisition. Nat. Methods, 17, 1229-1236. https://doi.org/10.1038/s41592-020-00998-0
- V. Demichev, C. Messner, S. Vernardis, K.S. Lilley, M. Ralser (2020). DIA-NN: neural networks and interference correction enable deep proteome coverage in high throughput. Nat. Methods, 17, 41-44. https://doi.org/10.1038/s41592-019-0638-x
- C. Lazar, L. Gatto, M. Ferro, C. Bruley, T. Burger (2016). Accounting for the multiple natures of missing values in label-free quantitative proteomics data sets to compare imputation strategies. J. Proteome Res., 15, 1116-1125. https://doi.org/10.1021/acs.jproteome.5b00981
- G. Rosenberger, S. Liu, J. Röst, H.L. Ludwig, C. Buil, B. Borchers, R. Aebersold, L. Collins (2017). Statistical control of peptide and protein error rates in large-scale targeted data-independent acquisition analyses. Nat. Commun., 8, 14224. https://doi.org/10.1038/ncomms14224
Support Documents
DIA-Based Quantitative Proteomics Service
Discover how DIA-based quantitative proteomics can support comprehensive protein profiling and comparative studies by downloading our service overview.
Protein Quantification for Deeper Discovery
Gain a clearer understanding of protein quantification approaches and their applications in studying protein abundance changes by downloading our detailed brochure.