



Sequential window acquisition of all theoretical fragment ion spectra (SWATH-MS) is a data-independent acquisition (DIA) schedule in which the mass spectrometer fragments precursors inside fixed m/z windows across each liquid chromatography (LC) cycle, then stores MS/MS traces for later extraction (Gillet et al. 2012). Protein quantification in SWATH projects is not read from a single spectrum display. It comes from extracted fragment chromatograms matched under false discovery rate (FDR) control after digests are acquired on separate injections in typical label-free discovery work. The guide below defines SWATH in project language, contrasts acquisition logic with data-dependent acquisition (DDA), and links mechanisms to workflow, parameters, limits, and fit. It is not a vendor tuning sheet, not a biomarker applications catalog, and not a substitute for a locked methods appendix.
SWATH names a window plan. DIA names the acquisition class. Wide windows, cycle time, and library strategy must be declared with matrix and contrast design before cohort scaling.
In MtoZ Biolabs offerings, SWATH-style DIA is discussed when open relative contrasts on unlabeled digests fit the cohort and platform scope supports windowed acquisition with documented extraction.
What SWATH Means Versus General DIA Language
Project records should separate three terms.
DIA means MS/MS is triggered on m/z intervals without intensity-based precursor picking during the run. Many precursors can fragment inside each interval per cycle.
SWATH-MS usually means sequential, contiguous windows that tile a declared precursor m/z range so the full range is sampled across cycles. Window width and count define cycle time and overlap risk.
Targeted data extraction means software matches stored fragments to expected transitions from spectral libraries or from library-free identification routes, then integrates peak groups per run (Gillet et al. 2012).
SWATH quantification is an extraction claim on predefined windows and contrasts, not a promise that every theoretical peptide is measured in one run.
Ion mobility-enhanced DIA adds a separation axis before windowed MS/MS (Mechanism of 4D-DIA Quantitative Proteomics). Mobility-free SWATH remains valid when pilots show adequate extraction without mobility gains.
DDA, DIA, and SWATH Acquisition Logic
Mechanisms differ at acquisition time.
In DDA, the instrument selects individual precursors by intensity for MS/MS in each cycle. Abundant ions dominate sampling. Low-abundance ions may be missed when cycle capacity is saturated.
In DIA, declared m/z windows fragment all ions in range per cycle. Sampling is window-driven, not intensity-ranked at acquisition.
In SWATH-style DIA, windows advance across the precursor range in a repeating schedule. Each peptide is sampled when its precursor m/z falls inside an active window during elution.
Window width trades fragment overlap against m/z coverage per cycle. Narrow windows increase cycles and can under-sample fast elution peaks. Wide windows increase co-fragmentation within the window.
Open discovery keeps protein lists unconstrained at acquisition. Closed lists may move to parallel reaction monitoring (PRM) after discovery (4D-DIA vs 4D-PRM: Which 4D Quantitative Approach Suits Your Project Better).
Operational handoffs appear in Workflow of 4D-DIA Quantitative Proteomics and SWATH Proteomics.
How Protein Quantification Emerges from SWATH MS/MS Traces
Acquisition stops at stored spectra. Quantification continues in software.
Peptide identification under FDR control assigns sequences to fragment groups. Library-based extraction uses prior fragment evidence. Library-free engines identify directly from DIA maps with interference correction when configured (Demichev et al. 2020).
Peak groups integrate to fragment intensities per run. Normalization aligns runs. Protein rollup follows a declared rule. Only the predefined contrast is tested statistically.
Overlapping fragments from co-eluting precursors in the same window create interference. Extractors apply scoring and filtering; they do not magically separate every overlap without evidence.
Missing values often track low abundance, interference, and prep loss, not only window count (Lazar et al. 2016).
Statistical control of peptide and protein error rates in large DIA analyses remains mandatory in extraction design (Rosenberger et al. 2017). Method comparison context appears in DIA Data Analysis Methods in Proteomics.
SWATH Principle Linked Workflow Spine
A practical sequence shows where definitions become project actions.
- Confirm matrix, contrast, and relative quantitative claim.
- Prepare matched digests to injection-ready loads on separate injections for label-free work.
- Pilot SWATH window width and cycle time on the matrix; lock LC and window plan.
- Acquire cohort under the locked schedule with agreed QC inserts.
- Extract under unified FDR, library provenance, and interference filters.
- Normalize, handle missing values, test predefined contrast only.
- Optional PRM on a short list when list status changes.
Steps 1 through 2 are prep outside SWATH naming. Steps 3 through 4 are windowed acquisition mechanisms. Steps 5 through 6 are quantification mechanisms.
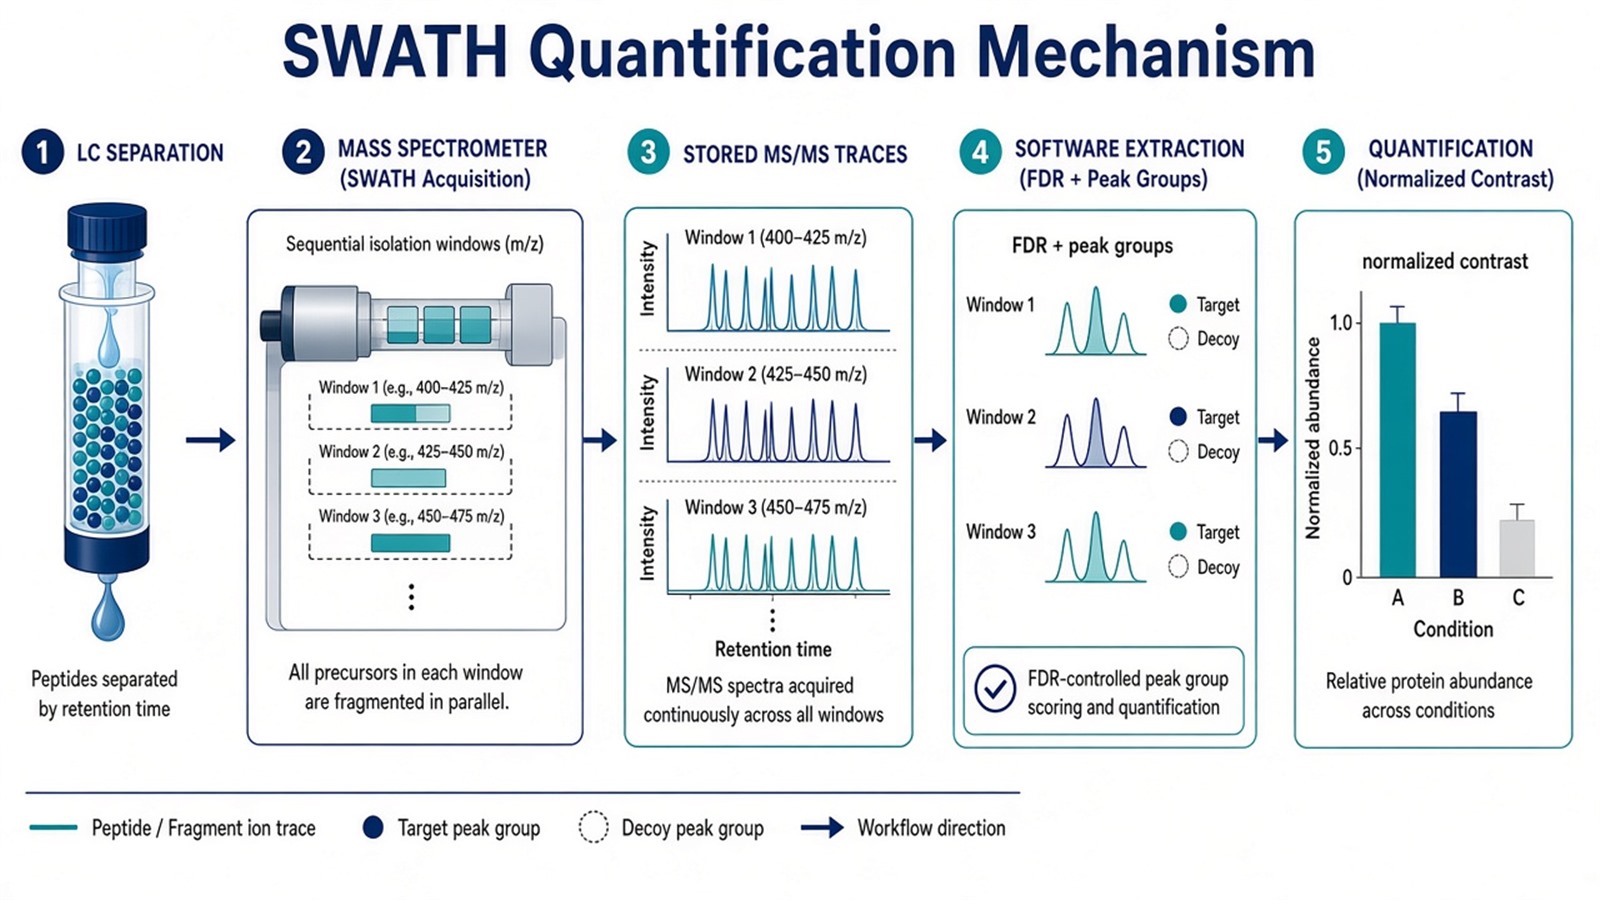
Figure 1. SWATH stores windowed MS/MS for extraction-based quantification; acquisition completeness is not the same as contrast-ready intensities.
Library workflow context appears in Quantitative Analysis of SWATH Proteomics Using OpenSWATH: A Complete Workflow Overview.
Parameters and Limits Tied to SWATH Quantification
Layer |
Parameter that must be declared |
Limit if mis-set |
|---|---|---|
LC |
Gradient length and peptide load |
Cycle under-sampling of elution peaks |
SWATH windows |
Width, count, m/z range tiled |
Overlap or coverage gaps |
Cycle time |
MS/MS duty versus elution width |
Missing fragments for fast peptides |
Library |
Source matrix and instrument generation |
Wrong expected transitions |
Extraction |
Library-based versus library-free route |
Biased search space |
FDR |
Unified model across files |
Drifting error control |
Contrast |
Frozen before threshold tuning |
Post hoc relabeling |
Claim |
Relative discovery default |
Open intensities treated as absolute amount |
Limits follow the same table. SWATH mechanisms do not by themselves prove diagnostic utility or drug mechanism. A digital archive of MS/MS does not replace matched pretreatment or frozen contrasts. Forced wide windows on sparse matrices may add overlap without depth gain.
Label-free chemistry is independent of SWATH scheduling (SWATH-MS Label-Free Protein Quantification Technology Analysis).
How to Choose SWATH-Based Quantification for a Project
Use pilot extraction quality, list status, and operations together.
Observation in pilots |
Mechanism interpretation |
Plan that often fits |
|---|---|---|
Stable extraction, open contrast |
Window plan adequate |
Label-free SWATH cohort |
High interference in wide windows |
Overlap dominates |
Narrower windows or pilot mobility DIA |
Closed peptide list after discovery |
Targeted remeasurement |
PRM on short list |
Amount on named peptides |
SIS targeted quant |
Not open SWATH intensities alone |
Simple matrix, clean DDA depth |
DIA may add prep cost only |
Compare DDA LFQ versus SWATH pilot |
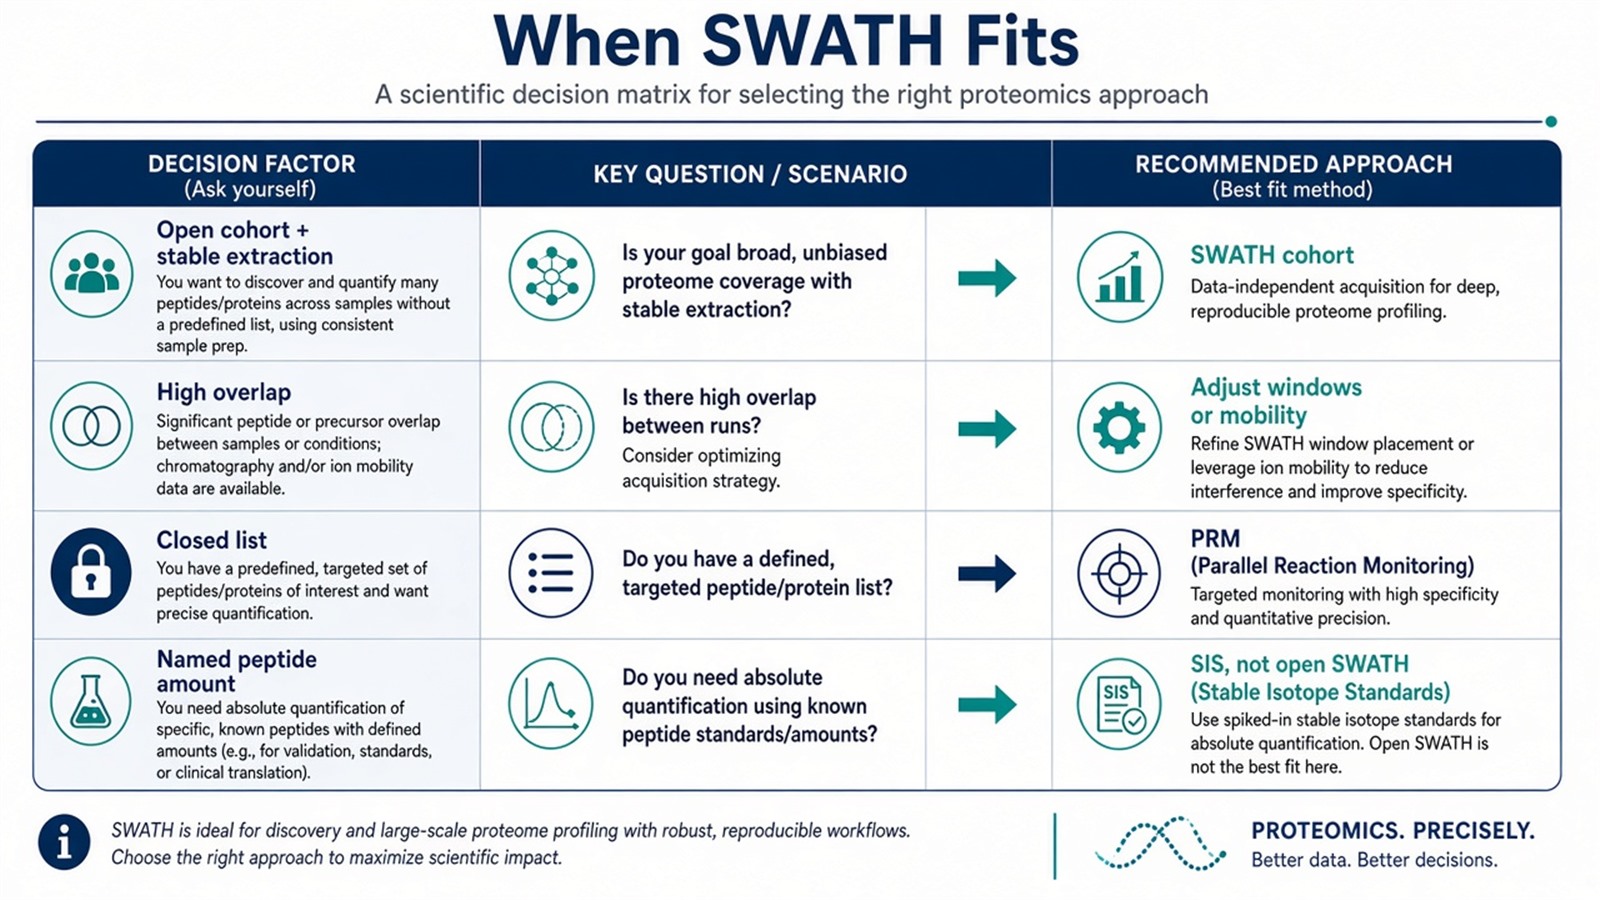
Figure 2. Choose SWATH when windowed DIA pilots deliver stable extraction for the frozen contrast; otherwise adjust windows, mobility, or move to PRM.
Three-axis route selection appears in Protein Quantification Method. Sample readiness checks appear in Exploring Quantitative Analysis in Proteomics.
Teams with pilot spectra, contrast design, and matrix documentation can review the MtoZ Biolabs Label-Free Quantitative Proteomics Service for SWATH mechanism and method alignment.
Frequently Asked Questions
1. Is SWATH the same as label-free quantification?
No. Label-free names unlabeled chemistry on separate injections. SWATH names a DIA window schedule. A project can use other DIA window plans without the SWATH label.
2. Does SWATH replace spectral libraries?
No. Libraries support targeted extraction in many SWATH pipelines. Library-free routes identify from DIA maps when declared in the analysis record.
3. Are 20 to 50 Da windows always correct?
No. Window width must fit elution density, instrument cycle time, and overlap tolerance on the matrix. Pilots lock values; generic ranges are not project defaults.
4. Can window settings change mid cohort?
Changing window programs mid study splits comparability. New settings need a new block and analysis record.
5. Does SWATH depth equal DDA identification lists?
No. DDA and DIA sample precursors differently. Depth and missing-value patterns must be judged against the predefined contrast, not against an unrelated DDA list from another run.
MtoZ Biolabs can discuss SWATH quantification principles in project planning when matrix pilots, contrast design, and extraction routes are documented.
If further evaluation of window design, library strategy, or discovery-to-targeted staging is needed, project details can be submitted for a feasibility review.
Reference
- L.C. Gillet, P. Navarro, S. Tate, H. Rost, N. Selevsek, L. Reiter, R. Bonner, R. Aebersold (2012). Targeted data extraction of the MS/MS spectra generated by data-independent acquisition: a new concept for consistent and accurate proteome analysis. Mol. Cell. Proteomics, 11, O111.016717. https://doi.org/10.1074/mcp.O111.016717
- V. Demichev, C. Messner, S. Vernardis, K.S. Lilley, M. Ralser (2020). DIA-NN: neural networks and interference correction enable deep proteome coverage in high throughput. Nat. Methods, 17, 41-44. https://doi.org/10.1038/s41592-019-0638-x
- C. Lazar, L. Gatto, M. Ferro, C. Bruley, T. Burger (2016). Accounting for the multiple natures of missing values in label-free quantitative proteomics data sets to compare imputation strategies. J. Proteome Res., 15, 1116-1125. https://doi.org/10.1021/acs.jproteome.5b00981
- G. Rosenberger, S. Liu, J. Röst, H.L. Ludwig, C. Buil, B. Borchers, R. Aebersold, L. Collins (2017). Statistical control of peptide and protein error rates in large-scale targeted data-independent acquisition analyses. Nat. Commun., 8, 14224. https://doi.org/10.1038/ncomms14224
- H.L. Röst, S. Liu, A. D'Agostino, M. Zanella, T. Navarro, P. Rosenberger, H. Collins, R. Aebersold (2017). OpenSWATH enables automated, targeted analysis of data-independent acquisition MS data. Nat. Biotechnol., 35, 219-223. https://doi.org/10.1038/nbt.3774
