



In targeted protein quantification, multiple reaction monitoring (MRM or SRM) schedules precursor-to-product transitions on a triple quadrupole: Q1 selects the peptide precursor, fragmentation occurs in the collision cell, and Q3 passes predefined product ions so only those transitions are recorded (Lange et al. 2008). Parallel reaction monitoring (PRM) on high-resolution instruments still isolates scheduled precursors but stores a full MS2 spectrum, allowing fragment chromatograms to be extracted and reviewed after acquisition (Peterson et al. 2012). Both mechanisms measure a closed peptide panel, not an open proteome. Default output is relative same-peptide signal; absolute amount needs stable isotope-labeled standards and calibration. Prefer MRM for mature high-multiplex transition panels; prefer PRM when accurate-mass fragment review helps complex digests.
Researchers who already have a closed peptide panel or a validation objective can also review the MtoZ Biolabs Parallel Reaction Monitoring (PRM) Service, SRM-MRM Quantitative Proteomics Services, and Targeted Proteomics Service for project-specific feasibility and assay planning.
Mechanism Comparison: MRM Versus PRM
Both routes answer the same class of question: how much signal does a predefined peptide surrogate produce under matched prep and scheduling. They differ in what the instrument records during acquisition.
Element |
MRM / SRM (typical triple quadrupole) |
PRM (typical high-resolution MS) |
|---|---|---|
Isolation |
Q1 precursor m/z |
Precursor isolation window |
Fragmentation |
Collision-induced dissociation in cell |
Similar fragmentation stage |
Detection |
Q3 filters predefined product transitions |
Full MS2 spectrum stored |
Quant feature |
Transition chromatogram peak areas |
Extracted fragment XICs after run |
Flexibility |
Transitions locked before acquisition |
Fragment choice refined post acquisition |
Multiplex driver |
Dwell time per transition |
Cycle time per scheduled precursor |
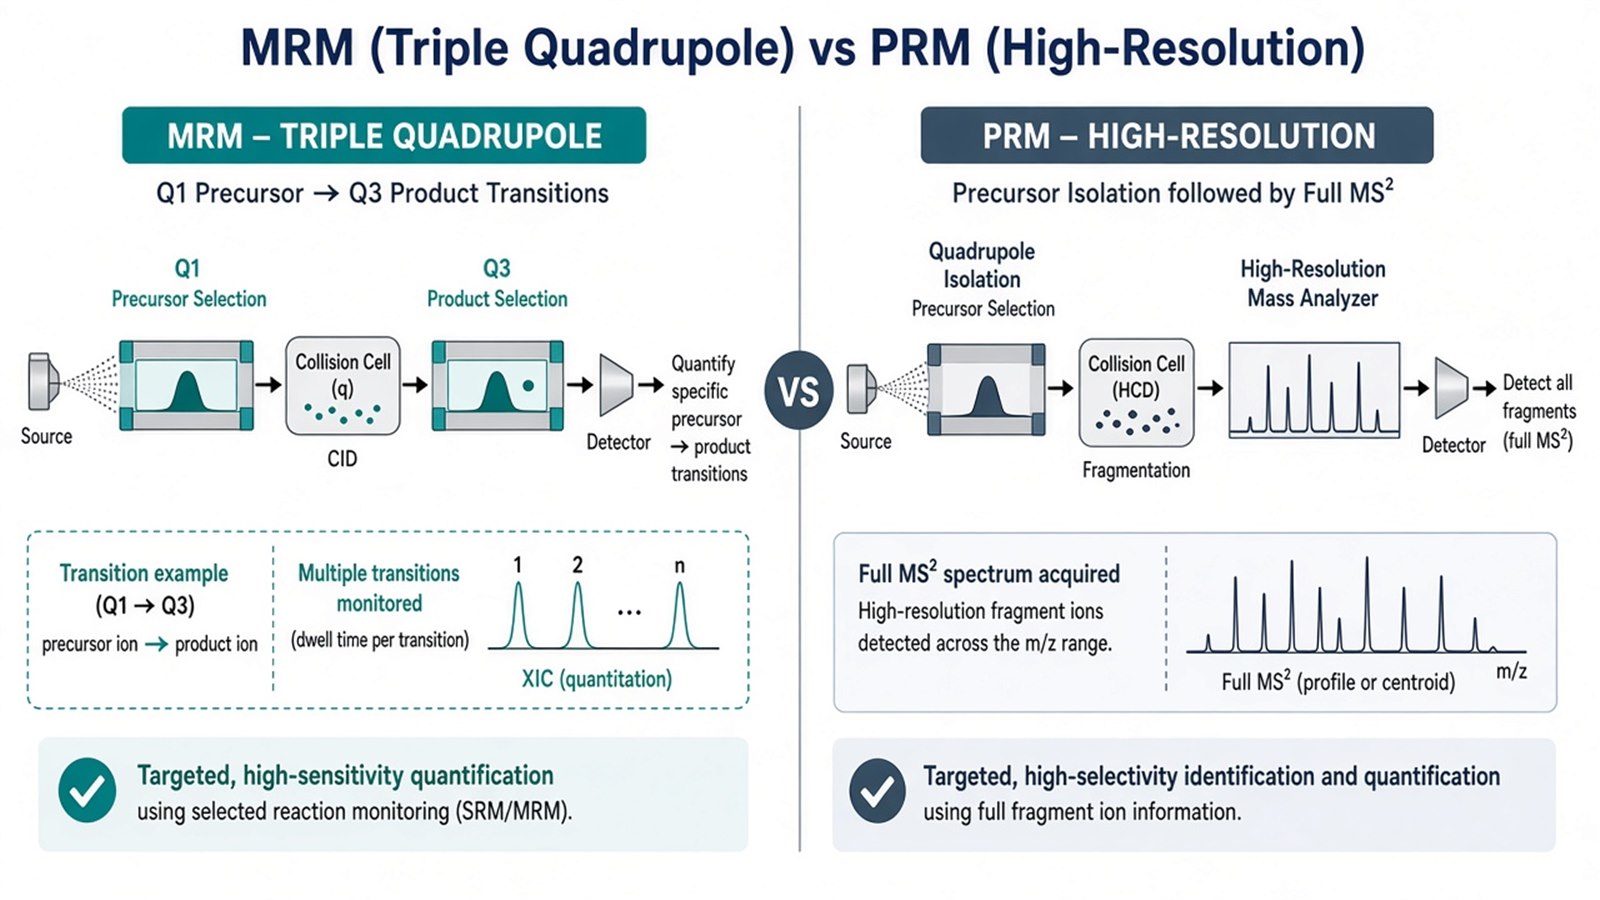
Figure 1. MRM quantifies predefined transitions during acquisition; PRM quantifies selected fragments from stored HR MS2 after acquisition.
PRM is not open discovery because many product ions are visible; only scheduled precursors enter the targeted quantification path.
For PRM-only principle depth, see Principles of PRM Quantitative Proteomics. For MRM principle context, see Principle of Multiple Reaction Monitoring and SRM Mass Spectrometry. For side-by-side selection, see PRM vs MRM: Comparative Analysis in Quantitative Proteomics.
Shared Targeted Quantification Spine
Mechanism sits inside a bottom-up workflow that is identical in prep requirements even when acquisition differs.
- Nominate proteotypic peptides from discovery or prior evidence.
- Match lysis, digestion, and load across contrast arms.
- Build inclusion lists or transition tables with charges and modifications declared.
- Schedule retention times and cycle time to preserve chromatographic peak sampling.
- Acquire MRM transitions or PRM HR MS2 for the closed panel only.
- Integrate peaks, check qualifiers, and normalize across runs.
- Optional SIS peptides and calibration curves when amount units are in scope.
- Report only inside the declared relative or absolute claim.
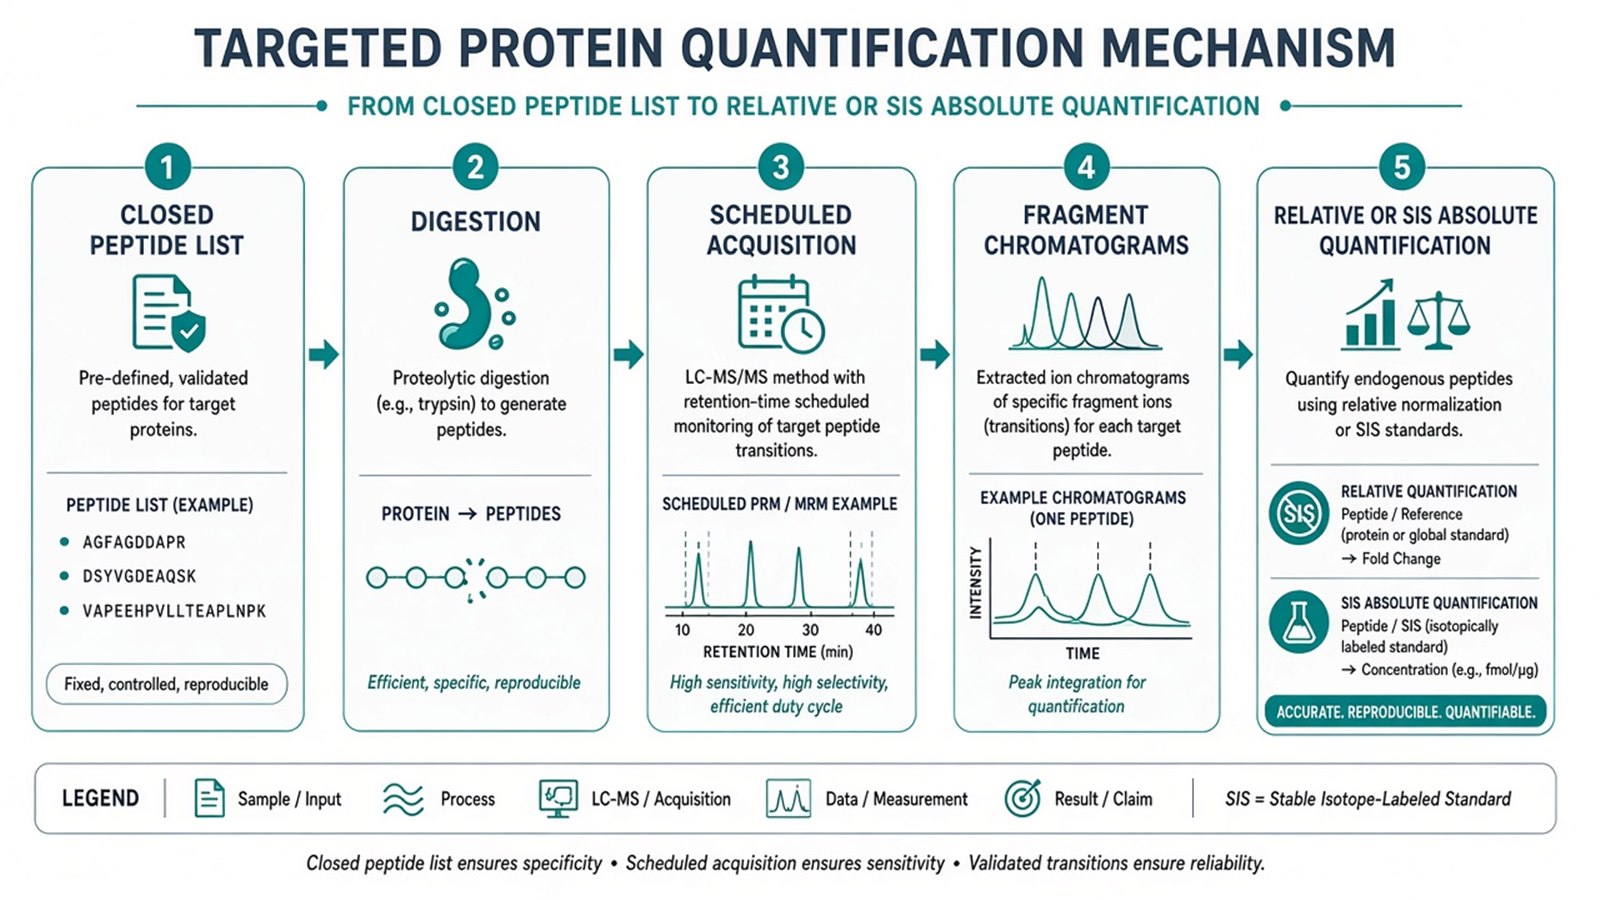
Figure 2. MRM and PRM mechanisms share closed-list targeting and matched prep; they diverge in how fragment ions are recorded and reviewed.
For PRM data operations, see PRM Proteomics Data Processing Workflow. For acquisition workflow, see Parallel Reaction Monitoring (PRM): Principles, Workflow, and Applications. For validation design, see How to Plan a Targeted Proteomics Validation Study.
Judgment Criteria: When MRM or PRM Mechanisms Fit
Study need |
MRM mechanism often fits |
PRM mechanism often fits |
Reassess |
|---|---|---|---|
Inventory |
Closed transitions on triple quadrupole |
Closed list needing HR MS2 review |
Candidates still expanding |
Matrix |
Clean panels with stable transitions |
Complex digests with interference risk |
Open discovery still required |
Multiplex |
High transition count with mature dwell plans |
Moderate list within HR cycle budget |
Overpacked schedule |
Claim |
Relative or SIS absolute on panel |
Relative or SIS absolute on panel |
Diagnostic cutoff without validation |
Upstream |
Discovery already nominated peptides |
Same |
No peptide evidence yet |
Prefer MRM when transitions are validated and triple-quadrupole multiplex is the throughput bottleneck. Prefer PRM when post-acquisition fragment review and accurate-mass specificity dominate. Prefer DIA or LFQ while the protein inventory must stay open. Prefer AQUA-style designs when concentration units require matrix-matched calibration, not only light-to-heavy ratios.
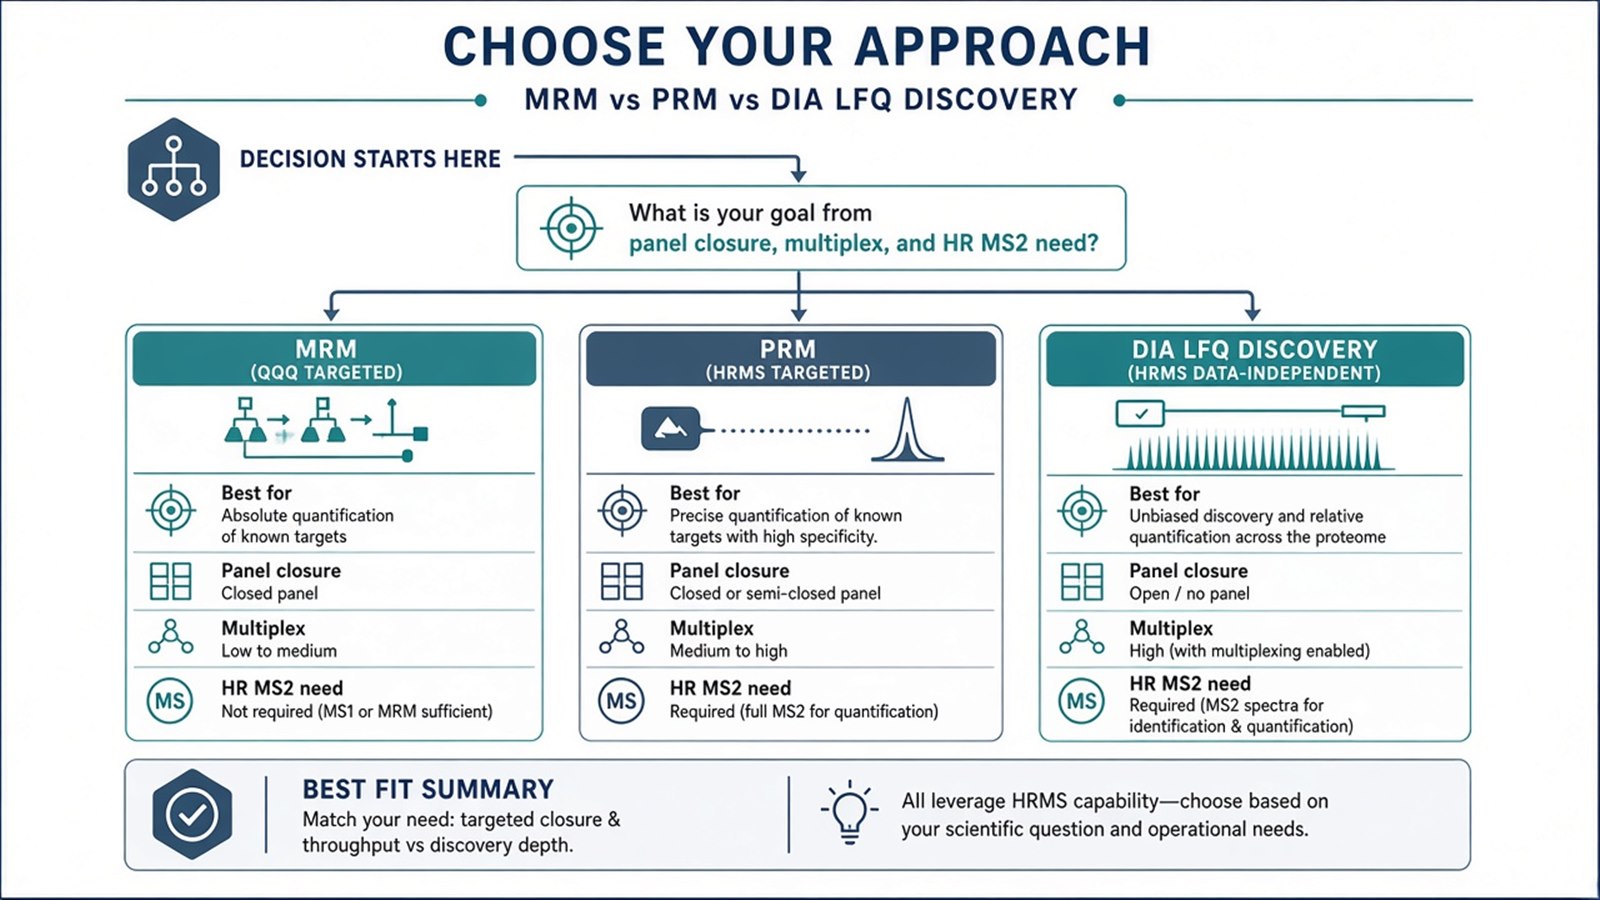
Figure 3. Choose MRM or PRM after the peptide list and claim are closed; discovery routes stay open until targeting is justified.
For advantage trade-offs on PRM, see Advantages and Disadvantages of Parallel Reaction Monitoring and PRM and MRM in Targeted Proteomics: Advantages and Limitations. For MRM applications, see Application of Multiple Reaction Monitoring (MRM) in Protein Quantification Analysis. For DIA handoff, see Application of DIA-PRM Proteomics.
Limits and Application Boundaries
Neither mechanism quantifies off-panel proteins. MRM unit-resolution transitions can suffer interference when matrices change. PRM co-isolation inside the window can still distort fragments if integration ignores qualifiers. Relative transition or fragment areas are not molar concentrations without declared SIS and calibration. Protein inference depends on surrogate peptides and digestion consistency. Biomarker discovery language must not be confused with diagnostic cutoffs unless a separately validated assay framework exists.
Typical applications when mechanism and prep are aligned:
- Confirmation of discovery-nominated biomarker peptides.
- Pharmacodynamic or signaling panels on predefined lists.
- Biopharmaceutical or comparability peptide monitoring.
- Absolute peptide amount on a shortlist when SIS and curves are planned.
- PRM follow-up when MRM transitions are unstable in complex digests.
Diagnostic cutoffs and companion-diagnostic claims sit outside ordinary research MRM or PRM reporting unless separately validated.
Once panel definition, mechanism choice, and claim type are fixed, the project can be scoped as MRM or PRM targeted quantification. Researchers can review the MtoZ Biolabs Parallel Reaction Monitoring (PRM) Service and SRM-MRM Quantitative Proteomics Services for sample evaluation and assay planning.
Frequently Asked Questions
1. What is the core mechanism shared by MRM and PRM?
Both isolate scheduled peptide precursors and quantify predefined surrogates in a closed panel under matched prep.
2. How does PRM differ mechanistically from MRM?
MRM records predefined Q1-Q3 transitions during acquisition. PRM stores full HR MS2 and extracts quantifier fragments after acquisition.
3. Can MRM or PRM quantify the whole proteome?
No. Only scheduled peptides or transitions are measured. Open coverage belongs to DIA or LFQ discovery routes.
4. When is absolute quantification part of the mechanism?
When SIS peptides and calibration logic are added to the same targeted panel. Otherwise the default mechanism output is relative.
5. Should discovery software process MRM or PRM data?
Targeted pipelines built for transition or PRM chromatograms fit these mechanisms. Discovery search engines alone do not replace targeted integration and QC rules.
Reference
- V. Lange, E. Picotti, B. Domon, R. Aebersold (2008). Selected reaction monitoring for quantitative proteomics: a tutorial. Mol. Syst. Biol., 4, 222. https://doi.org/10.1038/msb.2008.61
- A.C. Peterson, J.D. Russell, D.J. Bailey, M.S. Westphall, J.J. Coon (2012). Parallel reaction monitoring for high resolution and high mass accuracy quantitative, targeted proteomics. Mol. Cell. Proteomics, 11, 1475-1488. https://doi.org/10.1074/mcp.O112.020131
