



Label-free quantitative analysis of plant proteins usually means label-free quantification (LFQ) on unlabeled plant digests acquired as separate liquid chromatography-tandem mass spectrometry (LC-MS/MS) injections. Typical application questions compare relative protein change across treatments, tissues, genotypes, or time points after drought, salinity, heat, pathogen challenge, or developmental shifts (Bantscheff et al. 2012). In project language, plant LFQ names matched harvest and extraction, a frozen contrast, locked acquisition, and one processing chain. It is not a breeding outcome promise, not a substitute for genetics-only claims, and not an absolute amount assay without targeted follow-up (Gerber et al. 2003). The guide below maps plant LFQ applications to workflow, parameters, limits, and route choice. It is not a general plant biology textbook, not a vendor instrument catalog, and not a pathway enrichment manual.
Plant matrices add cell walls, starch, chloroplast-abundant proteins, and phenolic interference that animal or cell-line prep routes may not handle. Application value depends on prep parity across arms and on whether the contrast tests plant protein change, not confounded harvest batch (Cox et al. 2014).
In MtoZ Biolabs offerings, plant LFQ is evaluated when open contrasts, documented tissue handling, and unlabeled separate injections fit the cohort block.
General LFQ scope appears in LFQ. Application framing across domains appears in Label-Free Quantitative Proteomics: LFQ Technology, Applications, and Advances.
What Plant LFQ Applications Cover in Project Records
Application scope should name tissue, contrast, and claim unit together.
Growth and development applications compare organs, stages, or mutants when prep and harvest windows match across arms.
Abiotic stress applications compare control versus drought, salt, heat, or nutrient shift when stress starts and harvest times are documented.
Biotic stress applications compare mock versus inoculated material when infection stage and tissue zone are frozen in the brief.
Genotype-by-treatment applications compare cultivars or edited lines under the same environmental block when plot or chamber layout is recorded.
Plant LFQ applications deliver relative protein contrasts on declared groups. They do not by themselves validate field yield, variety release, or resistance breeding decisions.
Root-focused DIA context appears in SWATH Quantitative Proteomics of Plant Root Systems. Broader plant SWATH planning appears in SWATH-MS Enables Plant Proteomics Research: from Experiment to Data Analysis.
Plant Matrix and Contrast Patterns That Fit Label-Free LFQ
Plant applications fail early when matrix handling is underspecified.
Leaf, stem, root, seed, and pollen differ in Rubisco or storage protein dominance, wall material, and metabolite load. Subproteome goals may need compartment enrichment declared upfront.
Phenolic and polysaccharide carryover from insufficient cleanup can suppress ionization. Cleanup rules must match on every arm.
Harvest batch confounds watering, light, and clock time with treatment. Interleaved LC-MS/MS order and same-day harvest blocks reduce false stress signatures.
Open enrollment of greenhouse pots fits unlabeled separate injections. Closed same-day multiplex may fit isobaric tagging when all arms can be labeled together.
High missingness on low-abundance regulators in data-dependent acquisition (DDA) LFQ may push pilots toward unlabeled data-independent acquisition (DIA) when evidence supports it (Gillet et al. 2012).
Prep discipline appears in Quantitative Protein Analysis. LC-MS/MS scope appears in Label-Free LC-MS/MS.
End-to-End Workflow for Plant Label-Free LFQ
A practical plant LFQ application spine usually runs as follows.
- Define tissue, species, contrast, and relative claim before scaling harvest.
- Document growth chamber, field block, or hydroponic conditions tied to each arm.
- Harvest with matched timing; flash-freeze or stabilize with one declared protocol.
- Extract proteins with one plant-compatible route; remove debris and interfering metabolites equally on all arms.
- Digest and clean peptides to injection-ready load from a pilot when matrix is new.
- Acquire separate LC-MS/MS injections with interleaved order across groups when possible.
- Search with species-appropriate database, FDR, and modification rules.
- Extract LFQ features; normalize; apply missing-value policy; run QC before statistics.
- Test predefined contrast only; archive prep, methods, and software versions with deliverables.
Steps 1 through 5 are plant application layers. Steps 6 through 9 are shared LFQ layers.
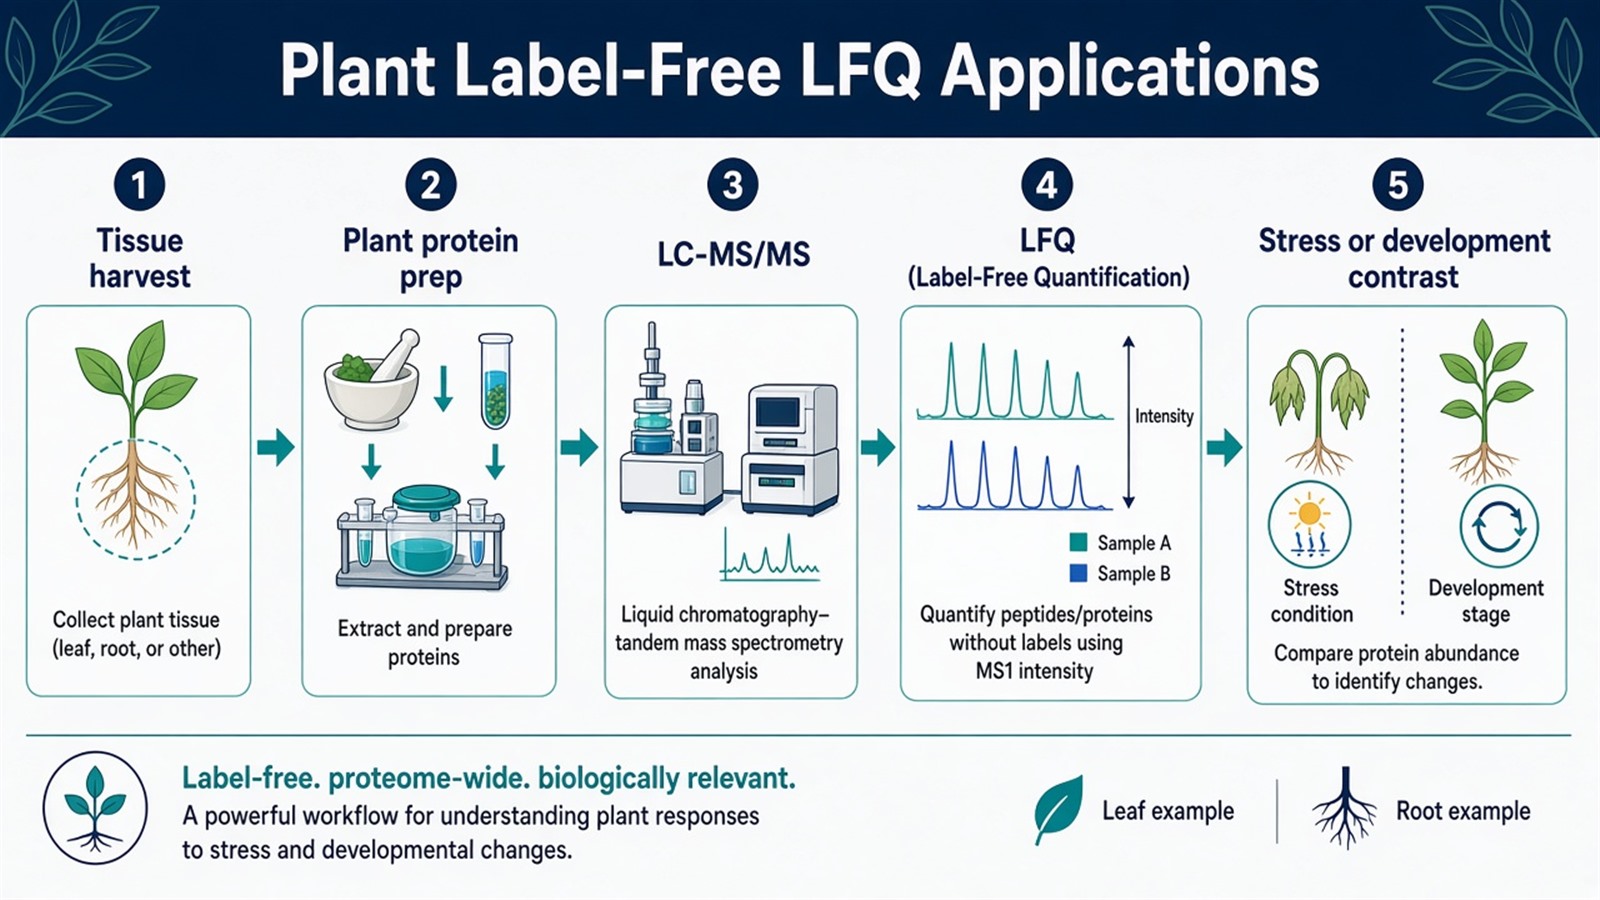
Figure 1. Plant LFQ applications depend on matched harvest and prep before acquisition; treatment effects must not confound with harvest batch.
Quality gates appear in How to Improve Data Quality and Reproducibility in Label-Free Proteomics?. Depth expectations appear in How Many Proteins Can Label-Free Detect?.
Parameters That Must Be Locked in Plant LFQ Runs
Layer |
Parameter that must be declared |
Failure mode if ignored |
|---|---|---|
Species |
Database and gene model version |
Wrong protein inference |
Tissue |
Organ, zone, developmental stage |
Mixed proteome slices |
Environment |
Block, chamber, or plot map |
Confounded treatment |
Harvest |
Clock time and handling |
Circadian and batch artifacts |
Extraction |
Buffer, ratio, cleanup |
Phenolic drift between arms |
Digest |
Enzyme and reduction rules |
Peptide pool shifts |
Load |
Mass on column from pilot |
Rubisco saturation |
Acquisition |
DDA or DIA template locked |
Mixed modes in one matrix |
Search |
FDR and modifications |
Inflated ID lists |
Contrast |
Frozen before tuning |
Post hoc relabeling |
Limits follow the same table. Plant LFQ does not quantify field performance, certify stress-tolerant varieties, or replace replicated agronomic trials. Abundant photosynthetic proteins can dominate MS time while regulators stay missing in DDA (Lazar et al. 2016). Rhizosphere or symbiosis studies may co-extract non-plant peptides unless washing and contrast scope are explicit.
Agricultural SWATH context appears in Applications and Breakthroughs of SWATH Proteomics in Agricultural Research. Normalization choices appear in How to Normalize Data in Label-Free Quantitative Proteomics?.
How to Choose Plant LFQ Versus Other Routes
Use application question, matrix pilot, enrollment, and follow-up unit together.
Plant application need |
Plan that often fits |
Recheck before committing |
|---|---|---|
Greenhouse stress time course |
DDA LFQ separate injections |
Harvest day tracks treatment |
High DDA missingness on targets |
Unlabeled DIA or SWATH-style LFQ |
DDA when pilot shows no gain |
Root–microbe contrast |
Matched wash; explicit claim unit |
Microbial co-extract as plant signal |
Multi-plot genotype study |
Interleaved runs; blocked harvest |
One genotype prep shortcut |
Closed arms, one labeling day |
TMT or iTRAQ multiplex |
Forced LFQ when tags required |
Seed or storage protein focus |
Declared tissue and depletion rule |
Depletion on one arm only |
Candidate proteins for breeding follow-up |
LFQ discovery then PRM |
Discovery list as validated marker |
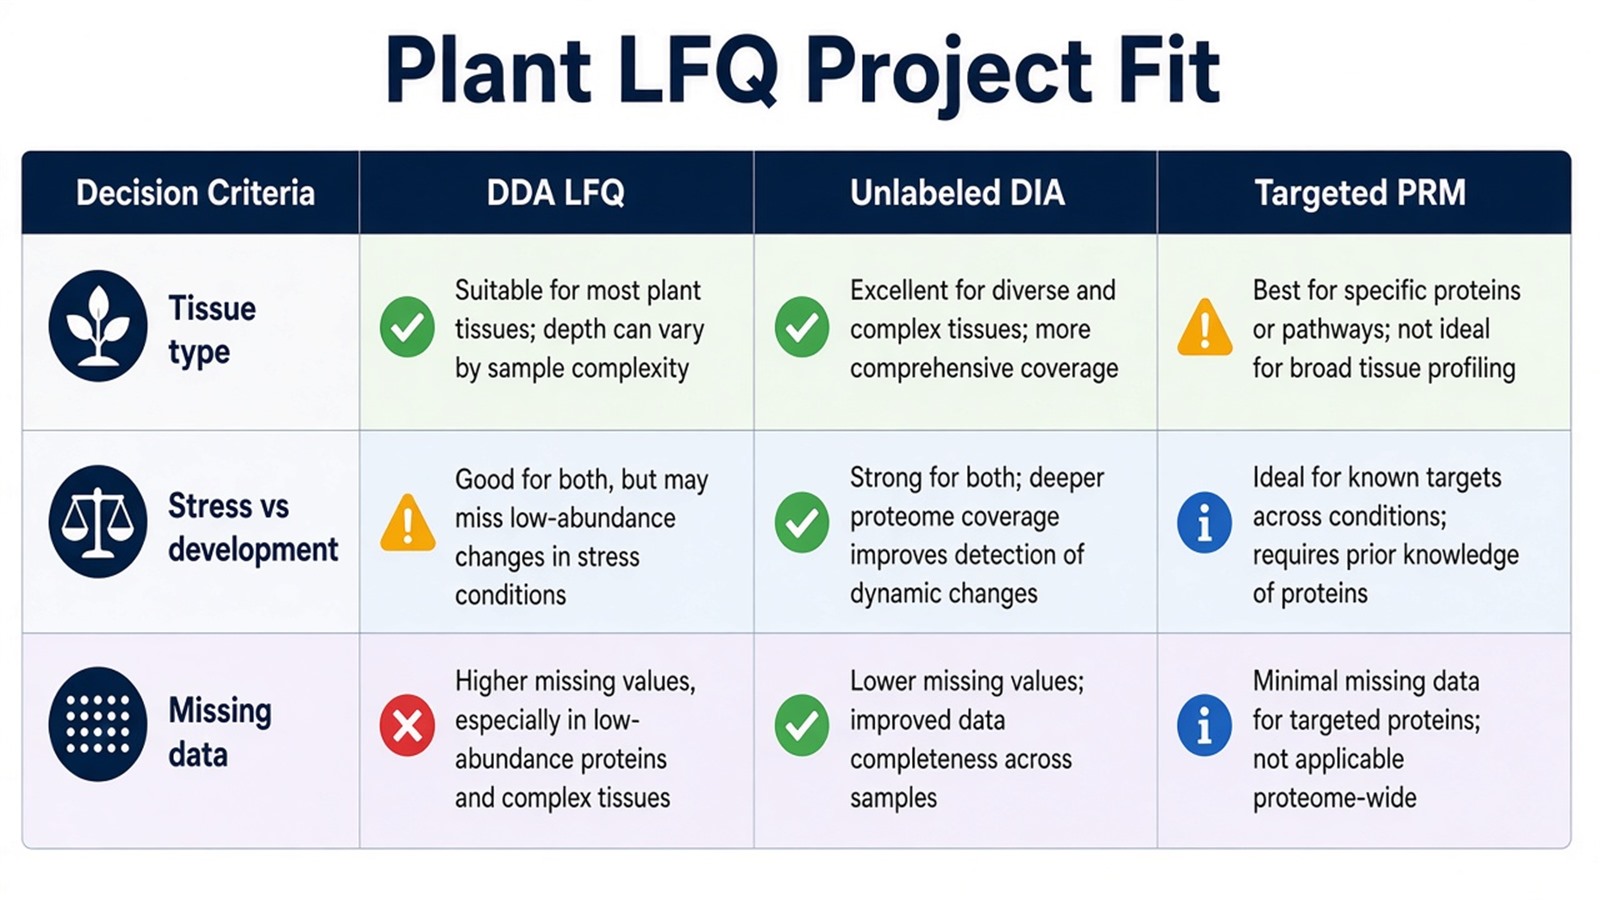
Figure 2. Plant LFQ fits open relative contrasts with matched prep; DIA, tagging, or targeted routes fit other missingness and claim patterns.
Route selection appears in Protein Quantification Method. Label-free tradeoffs appear in Advantages of Label-Free Proteomics Analysis.
Teams with species, tissue map, contrast design, and pilot notes can review the MtoZ Biolabs Label-Free Quantitative Proteomics Service for plant LFQ application alignment when declared in the project record.
Frequently Asked Questions
1. Which plant tissues work for label-free LFQ?
Many tissues work when extraction and cleanup are matched. Leaf, root, stem, flower, and seed differ in prep burden and abundant protein background. Pilot runs should set load and depth expectations per tissue.
2. Can plant LFQ replace transcriptomics for stress mechanism claims?
LFQ reports relative protein change on tested contrasts. Transcript and protein units differ. Integration is interpretive and should not mix undefined contrasts.
3. Should Rubisco depletion be used?
Depletion or fractionation is a project decision. If used, it must apply equally to every arm in the contrast and be recorded in the methods appendix.
4. Is label-free LFQ enough for disease-resistance breeding markers?
LFQ can prioritize candidate proteins for follow-up. Breeding markers need separate validation design, not discovery contrasts alone.
5. When should plant projects move from DDA LFQ to unlabeled DIA?
When pilots show acceptable identification but poor quantifiable coverage on proteins near the application decision threshold, unlabeled DIA pilots are a common next step if window schedules fit the matrix.
MtoZ Biolabs can discuss plant LFQ application planning when species, tissue handling, contrast map, and follow-up intent are documented.
If further evaluation of plant prep pilots, DDA versus unlabeled DIA fit, or discovery-to-targeted staging is needed, project details can be submitted for a feasibility review.
Reference
- M. Bantscheff, S. Lemeer, H.U. Haab, B. Kuster (2012). Quantitative mass spectrometry in proteomics: a critical review. Anal. Bioanal. Chem., 404, 939-965. https://doi.org/10.1007/s00216-012-5892-4
- J. Cox, M.Y. Hein, C.A. Luber, I. Paron, M. Mann (2014). Accurate proteome-wide label-free quantification by delayed normalization and maximal peptide ratio extraction, termed MaxLFQ. Mol. Cell. Proteomics, 13, 2513-2526. https://doi.org/10.1074/mcp.M113.031031
- L.C. Gillet, P. Navarro, S. Tate, H. Rost, N. Selevsek, L. Reiter, R. Bonner, R. Aebersold (2012). Targeted data extraction of the MS/MS spectra generated by data-independent acquisition: a new concept for consistent and accurate proteome analysis. Mol. Cell. Proteomics, 11, O111.016717. https://doi.org/10.1074/mcp.O111.016717
- C. Lazar, L. Gatto, M. Ferro, C. Bruley, T. Burger (2016). Accounting for the multiple natures of missing values in label-free quantitative proteomics data sets to compare imputation strategies. J. Proteome Res., 15, 1116-1125. https://doi.org/10.1021/acs.jproteome.5b00981
- S.A. Gerber, J. Rush, O. Stemman, M.W. Kirschner, S.P. Gygi (2003). Absolute quantification of proteins and phosphoproteins from cell lysates by tandem MS. Proc. Natl. Acad. Sci. U.S.A., 100, 6940-6945. https://doi.org/10.1073/pnas.0832254100
