



Optimize tandem mass tag (TMT) experimental design and sample prep before any liquid chromatography-tandem mass spectrometry (LC-MS/MS) queue: freeze a closed channel map that fits one commercial plex or a bridged multi-plex plan, match lysis and digestion across groups, normalize protein load before labeling, and remove residual amine buffers that compete with NHS-style tags. Incomplete labeling, unmatched loads, and scrambled channel maps become permanent ratio artifacts after pooling, and later MS2 or MS3 acquisition cannot repair them (Thompson et al. 2003; Ting et al. 2011). Interleave biological replicates and conditions across channels rather than stacking all controls together. TMT prep supports relative ratios inside that plex, not absolute concentration without matched standards.
Researchers who already have defined samples, comparison groups, or a specific project objective can also review the MtoZ Biolabs TMT Quantitative Proteomics Analysis Service for project-specific feasibility and sample planning.
Sample Prep Controls That Decide TMT Success
Relative TMT comparison assumes every channel entered the pool under comparable chemistry. Prep optimization therefore focuses on what happens before mixing.
Prep control |
Why it matters |
Failure mode if skipped |
|---|---|---|
Matched lysis and digestion |
Same recipe keeps recovery comparable across groups |
Protocol drift looks like biology after pooling |
Protein assay and equal loading |
Channel ratios reflect biology, not input mass |
One channel globally high or low |
Amine-buffer cleanup |
Free amines compete with TMT reagents |
Incomplete labeling and distorted reporters |
Quench and free-tag removal |
Excess reagent should not dominate spectra |
Chemical noise and labeling artifacts |
Locked channel map before pool |
Sample IDs must match reporter channels |
Swapped maps that normalization cannot fix |
Digestion QC before labeling |
Incomplete digests waste channels |
Sparse peptides and unstable ratios |
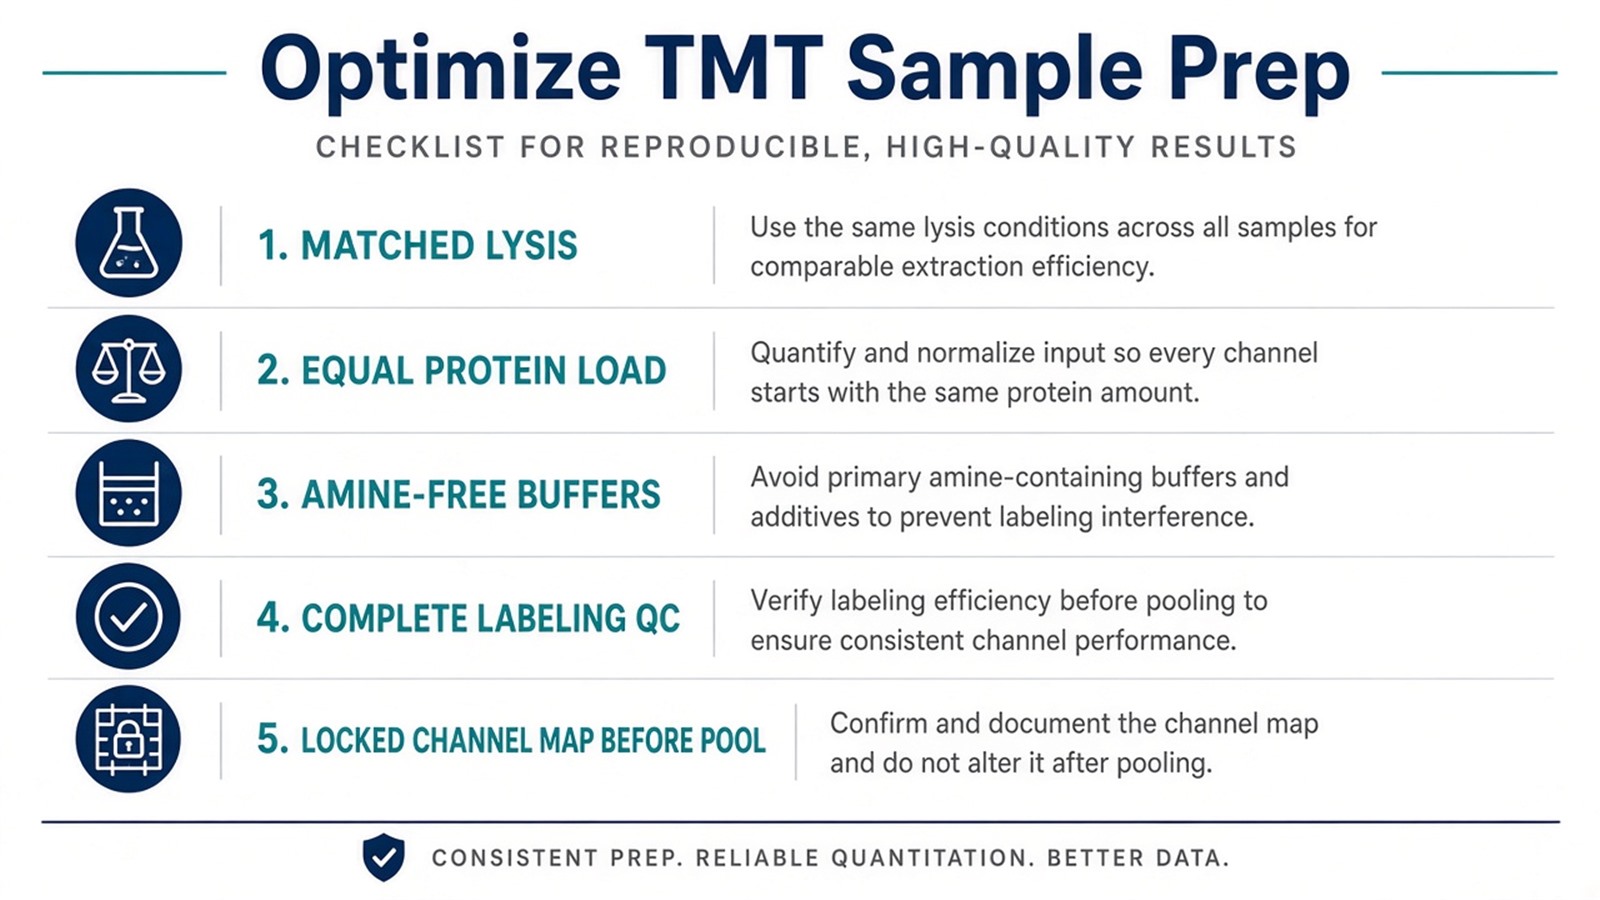
Figure 1. TMT sample-prep optimization is upstream of acquisition: matched digestion, amine-compatible chemistry, and a locked channel map come before pooling.
Prep failures become quantitative artifacts. Software cannot reconstruct protein that was lost or mislabeled before the plex was mixed.
Experimental Design Layout Before Labeling
Design optimization is about which samples share one plex and how channels are assigned.
1. Close enrollment for the plex
Decide which groups, time points, and biological replicates must be compared together. If samples will keep arriving, prefer label-free quantification (LFQ) or plan separate bridged plexes rather than forcing unrelated leftovers into spare tags.
2. Balance channels
Distribute conditions and replicates across the kit so one biological group is not trapped in adjacent channels or one instrument block. Randomize or block assignment under a written map before reagents are opened.
3. Plan bridges for multi-plex studies
When sample count exceeds one kit, include common reference or bridge channels that appear in every plex. Do not casually merge unbridged reporter matrices after the fact.
4. Reserve material for QC
Keep aliquots for labeling-efficiency checks, pooled QC digests, and optional fractionation tests. Absolute AQUA follow-up later still needs reserved material and a closed peptide list.
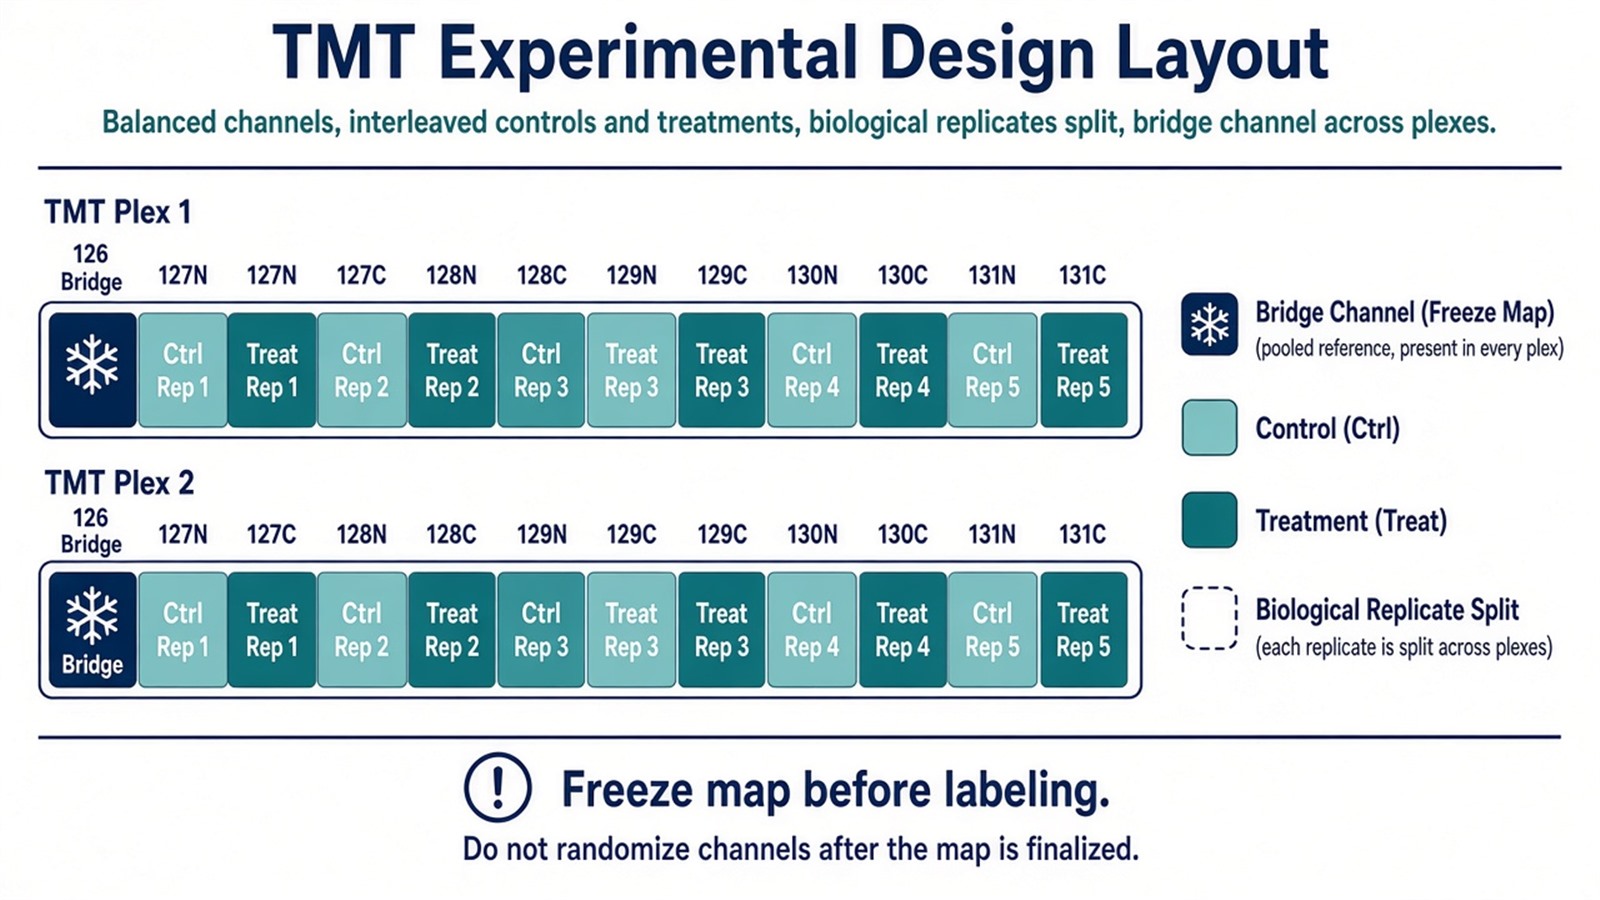
Figure 2. Optimize design by freezing a balanced channel map and bridging plan before labeling chemistry starts.
For operational labeling-to-acquisition steps after prep is locked, see TMT-Based Quantitative Proteomics: Multiplexing, MS3 Accuracy, and Workflow Design and A Detailed Workflow of TMT-Based Quantitative Proteomics. For labeling chemistry limits, see Mechanism of TMT Labeling in Mass Spectrometry. For ratio compression after imperfect prep, see How Does Ratio Compression Arise in TMT-Based Quantitative Proteomics and Why Is It Limited to Relative Quantification?.
Judgment Criteria: When Optimized TMT Prep Fits
Study need |
Optimized TMT design and prep often fit |
Reassess before labeling |
|---|---|---|
Cohort structure |
Groups and replicates fit one kit or a bridged plan |
Enrollment still open with no bridge design |
Chemistry |
Amine cleanup and complete labeling are feasible |
Residual Tris or competing amines dominate |
Input budget |
Enough material for matched loads and QC |
Only one scarce aliquot remains |
Claim |
Relative same-run ratios inside the plex |
Absolute concentration or diagnostic cutoff |
Alternative |
Same-run multiplex worth the prep cost |
Independent LFQ or DIA fits better |
Prefer optimized TMT prep when the cohort can be frozen and amine chemistry is clean. Prefer LFQ or unlabeled DIA when samples must stay independent or labeling would waste scarce material. Prefer SPS-MS3 and fractionation after prep is sound, not as a substitute for unmatched loads. Prefer PRM, MRM, or AQUA when targets are closed or amount units are required.
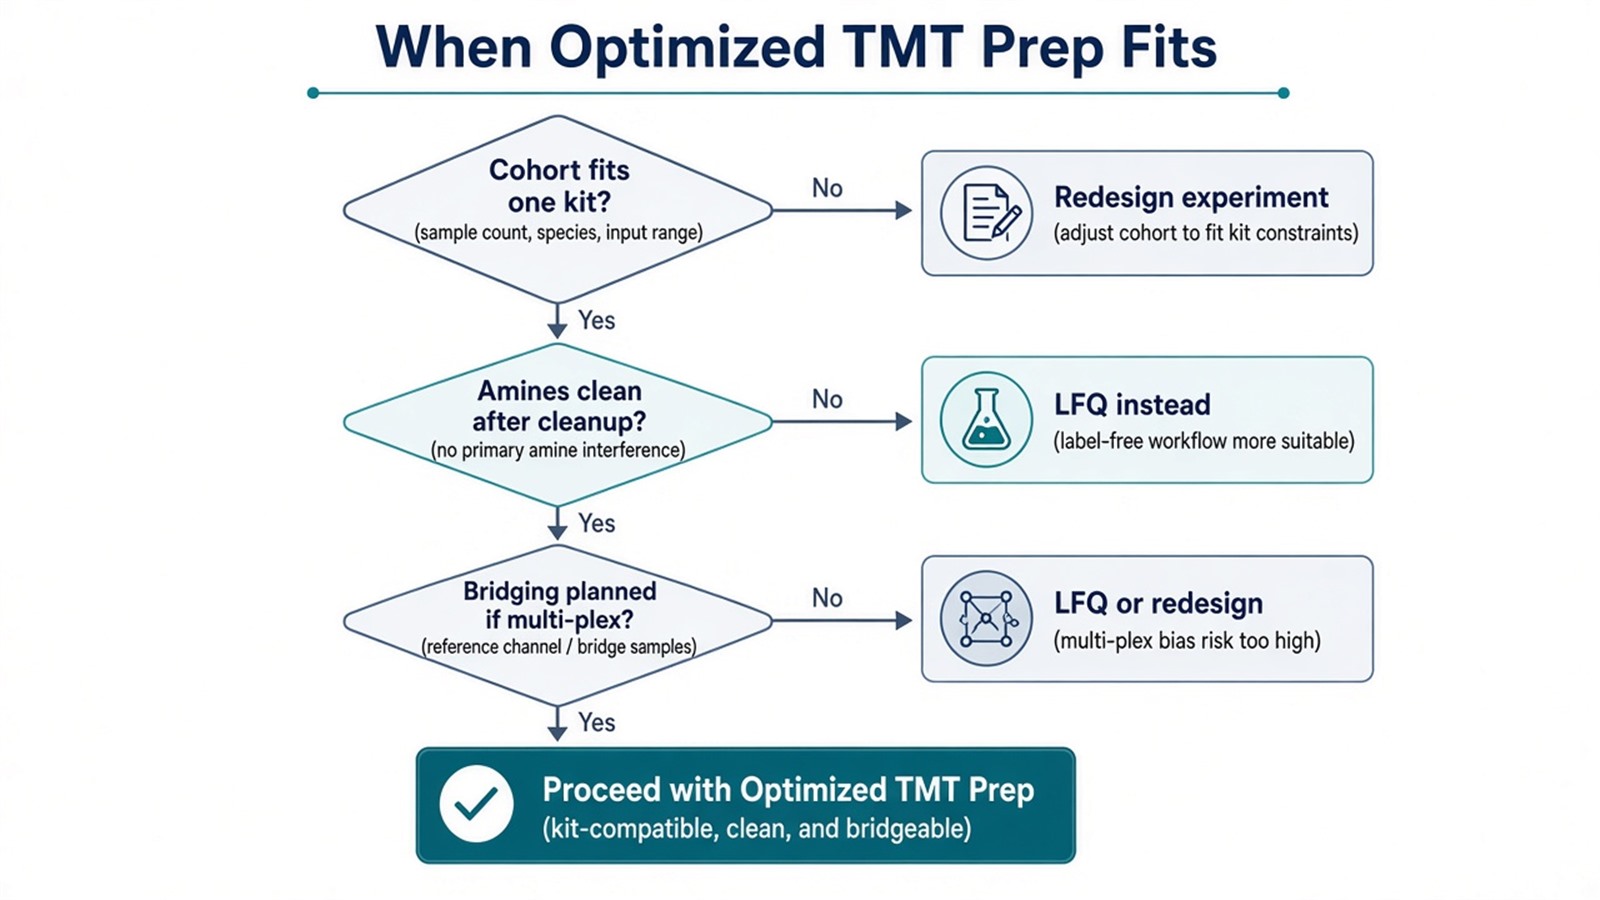
Figure 3. If plex closure, amine chemistry, or bridging fails, change the quantitative plan rather than forcing incompatible sample prep.
For selection against unlabeled routes, see TMT vs Label Free Quantitative Proteomics and Comparing LFQ and Isobaric Labeling Strategies in Proteomics Quantification. For sample constraints across absolute and relative claims, see Absolute and Relative Quantitative Proteomics Analysis. For SPS-MS3 after prep is locked, see How Can SPS-MS3 Be Utilized to Optimize TMT Workflows and Minimize Ratio Compression?.
Limits and Application Boundaries
Optimized prep does not convert TMT into absolute quantification. It does not erase co-isolation without fractionation or MS3 where appropriate. Pathway enrichment from a differential list still reflects overrepresentation relative to the quantified background, not proven pathway activity.
Typical applications that benefit from this design and prep focus:
- Multi-arm drug or genotype panels that can be labeled together under one kit.
- Time-course sets that can freeze all planned points before mixing.
- Tissue or cell cohorts where matched lysis and equal loading are enforceable.
- Multi-plex clinical or preclinical series only when bridge channels are planned in advance.
Diagnostic cutoffs and regulatory assay claims sit outside ordinary research TMT reporting unless a separately validated framework is in place.
Once sample type, buffer compatibility, channel map, and bridging rules are defined, the project can be scoped as a TMT study. Researchers can review the MtoZ Biolabs TMT Quantitative Proteomics Analysis Service for sample evaluation, feasibility assessment, and workflow planning. For design trade-offs, see Advantages and Disadvantages of TMT-Based Quantitative Proteomics Analysis.
Frequently Asked Questions
1. What sample-prep step most often breaks TMT ratios?
Unmatched protein loads and residual amine buffers are common. Both become channel artifacts after pooling.
2. Should all controls occupy neighboring TMT channels?
No. Balance conditions and replicates across the plex under a written map before labeling.
3. Can I add late samples to an already pooled plex?
No. Enrollment closes at mixing. Plan a new bridged plex or keep unlabeled LFQ if arrival is open-ended.
4. Does better sample prep remove the need for MS3?
Not always. Clean prep fixes chemistry bias. Co-isolation in complex digests may still need fractionation or SPS-MS3.
5. Is optimized TMT prep enough for absolute protein amount?
No. Absolute units need matched standards and calibration outside ordinary TMT discovery tables.
Reference
- A. Thompson, J. Schaefer, K. Kuhn, S. Kienle, J. Schwarz, G. Schmidt, T. Neumann, C. Hamon (2003). Tandem mass tags: a novel quantification strategy for comparative analysis of complex protein mixtures by MS/MS. Anal. Chem., 75, 1895-1904. https://doi.org/10.1021/ac0262560
- L. Ting, R. Rad, S.P. Gygi, W. Haas (2011). MS3 eliminates ratio distortion in isobaric multiplexed quantitative proteomics. Nat. Methods, 8, 937-940. https://doi.org/10.1038/nmeth.1714
- G.C. McAlister, D.P. Nusinow, M.P. Jedrychowski, M. Wuhr, E.L. Huttlin, B.K. Erickson, R. Rad, W. Haas, S.P. Gygi (2014). MultiNotch MS3 enables accurate, sensitive, and multiplexed detection of differential expression across cancer cell line proteomes. Anal. Chem., 86, 7150-7158. https://doi.org/10.1021/ac502040v
