



Quantitative proteomics measures protein abundance after bottom-up digestion and LC-MS/MS under a declared method family and claim. Relative quantification compares the same protein or peptide across matched samples; absolute quantification needs stable isotope standards on a closed panel (Cox et al. 2014; Thompson et al. 2003; Peterson et al. 2012). Families are unlabeled LFQ or DIA, isobaric TMT or iTRAQ, and targeted PRM, MRM, or AQUA. Shared workflow is prep, digestion, acquisition, and QC; branches differ by labeling, DDA versus DIA versus targeted schedules, and relative versus amount reporting. Prefer LFQ or DIA for open discovery; TMT for a frozen plex; targeted MS for confirmation or calibrated amount.
Researchers who already have defined samples, comparison groups, or a specific project objective can also review the MtoZ Biolabs Label-Free Quantitative Proteomics Service, TMT Quantitative Proteomics Analysis Service, and Targeted Proteomics Service for project-specific feasibility and analysis planning.
Methods Map for Quantitative Proteomics
LC-MS/MS is the common measurement layer. The method family defines chemistry, acquisition, and the defensible quantitative statement.
Method family |
Typical measurement |
Quantitative claim |
|---|---|---|
Label-free LFQ (DDA) |
Independent runs; MS1 peak areas |
Relative same-protein change |
Label-free DIA / SWATH-style |
Windowed MS/MS; fragment peak groups |
Relative discovery matrices |
Isobaric TMT / iTRAQ |
Pooled multiplex; reporter ions |
Relative ratios inside a plex |
Targeted PRM / MRM |
Scheduled peptides or transitions |
Relative or calibrated panel |
AQUA / SIS absolute |
Light-heavy pairs with curves |
Peptide amount under stated design |
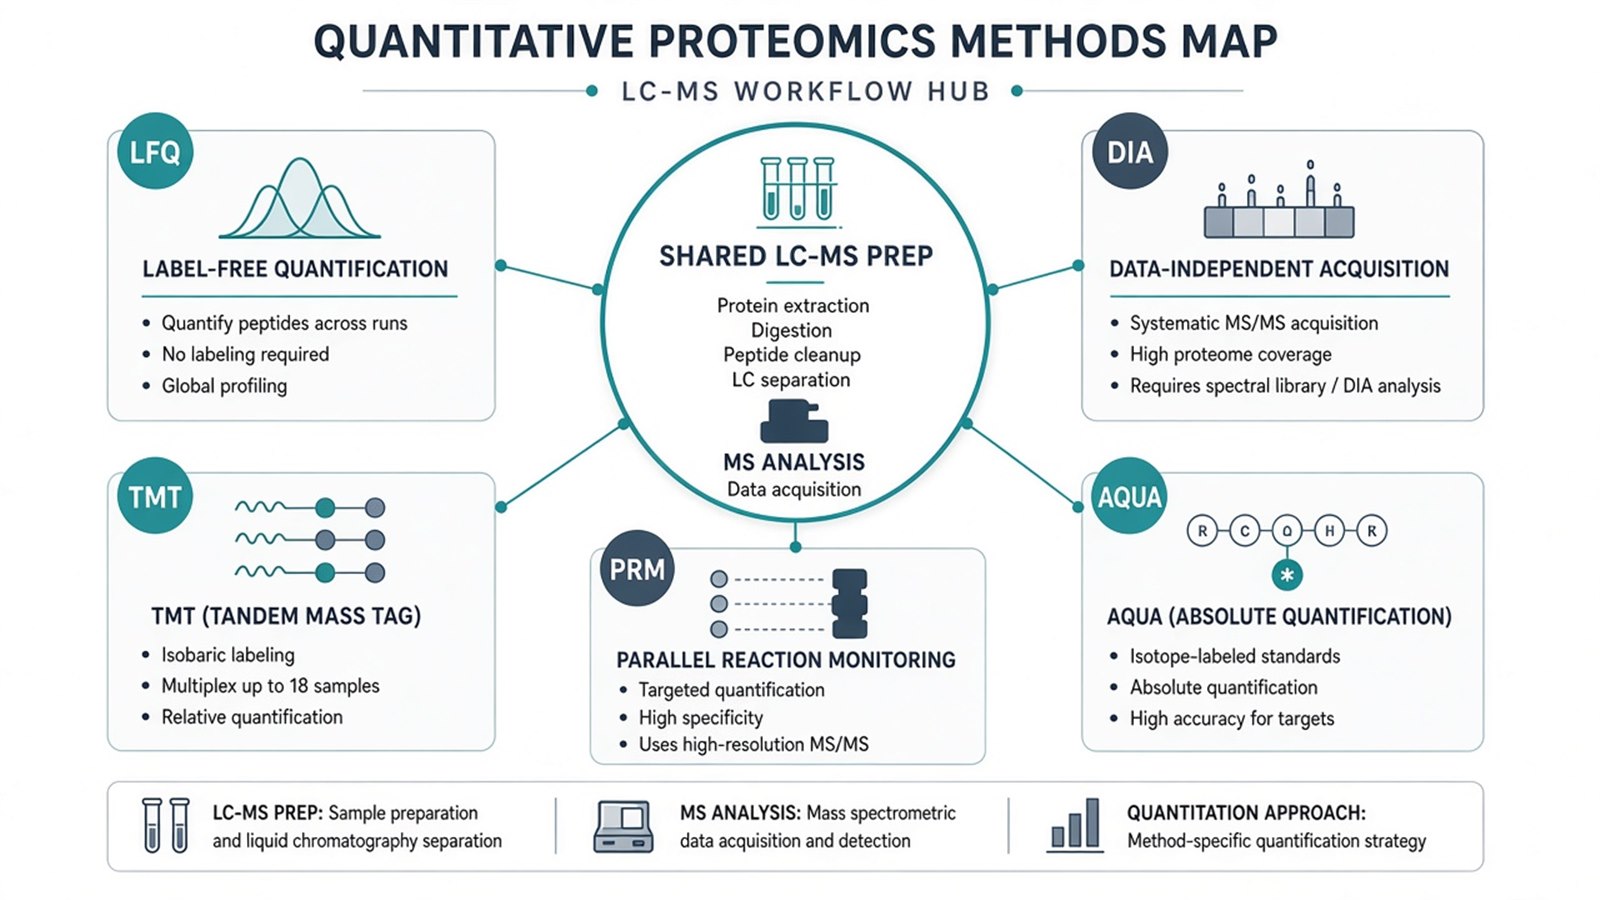
Figure 1. To quantify proteins by MS, name the method family and claim type; LC-MS alone is not a complete method name.
Equating protein quantification with any peak intensity table without stating relative versus absolute design leaves results underspecified.
For LC-MS framing detail, see LC-MS Protein Quantification. For labeled versus unlabeled selection, see Differences of Labeled and Label-free Quantitative Proteomics. For isotopic labeling overview, see Quantitative Analysis of Proteomics Using Isotopic Labeling Methods.
Shared Workflow From Sample to Quantitative Matrix
- Define contrasts, replicates, and relative or absolute intent before prep scales.
- Extract and digest proteins with matched chemistry across groups (Wisniewski et al. 2009).
- Branch: unlabeled independent injections; isobaric labeling and pool; or SIS spike for absolute targeted work.
- Separate peptides by LC; acquire DDA, DIA, or targeted schedules under locked methods.
- Control FDR or transition QC; normalize; export a protein-by-sample matrix.
- Test predefined contrasts; optional PRM, MRM, or AQUA on nominated peptides.
- Treat pathway enrichment as secondary interpretation, not a substitute for QC.
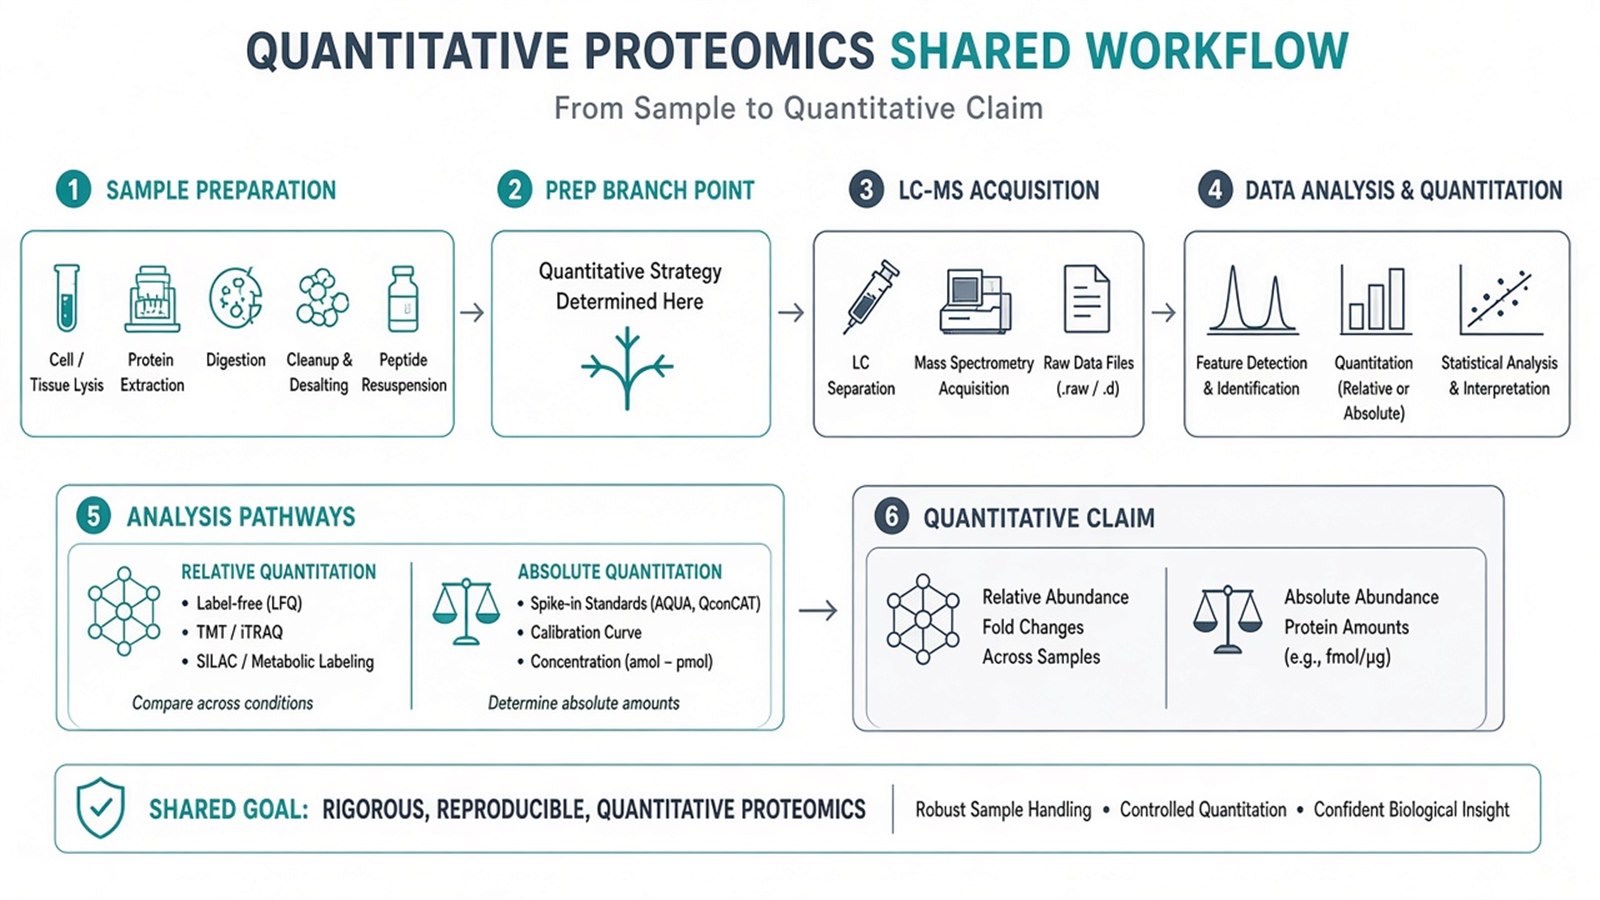
Figure 2. Workflow and applications connect at shared prep; they diverge at labeling, acquisition mode, and claim type.
For label-free experimental overview, see Overview of Label-Free Proteomics Experimental Workflow and How to Perform Label-Free Quantitative Proteomics Analysis Using Mass Spectrometry?. For DIA workflow, see DIA Proteomics Workflow: Acquisition, Library Strategy and Data Analysis. For absolute versus relative gates, see Absolute and Relative Quantitative Proteomics Analysis.
Judgment Criteria: Which Quantitative Route Fits
Study need |
Often prefer |
Reassess |
|---|---|---|
Open discovery; flexible enrollment |
LFQ or unlabeled DIA |
Absolute amount without standards |
Multi-arm panel in one multiplex mix |
TMT or iTRAQ |
Late samples after pooling |
Closed candidate list |
PRM or MRM |
Inventory still expanding |
Calibrated peptide amount |
AQUA / SIS |
No matrix-matched calibrators |
Prep-limited tough or low-input tissue |
Optional PCT then DIA |
Forcing labeling on scarce input |
Prefer unlabeled discovery when independent runs and matched prep fit the question. Prefer isobaric labeling when plex closure and amine-compatible prep hold. Prefer targeted assays when confirmation or amount units dominate. Prefer revisiting prep when ratios track handling rather than biology.
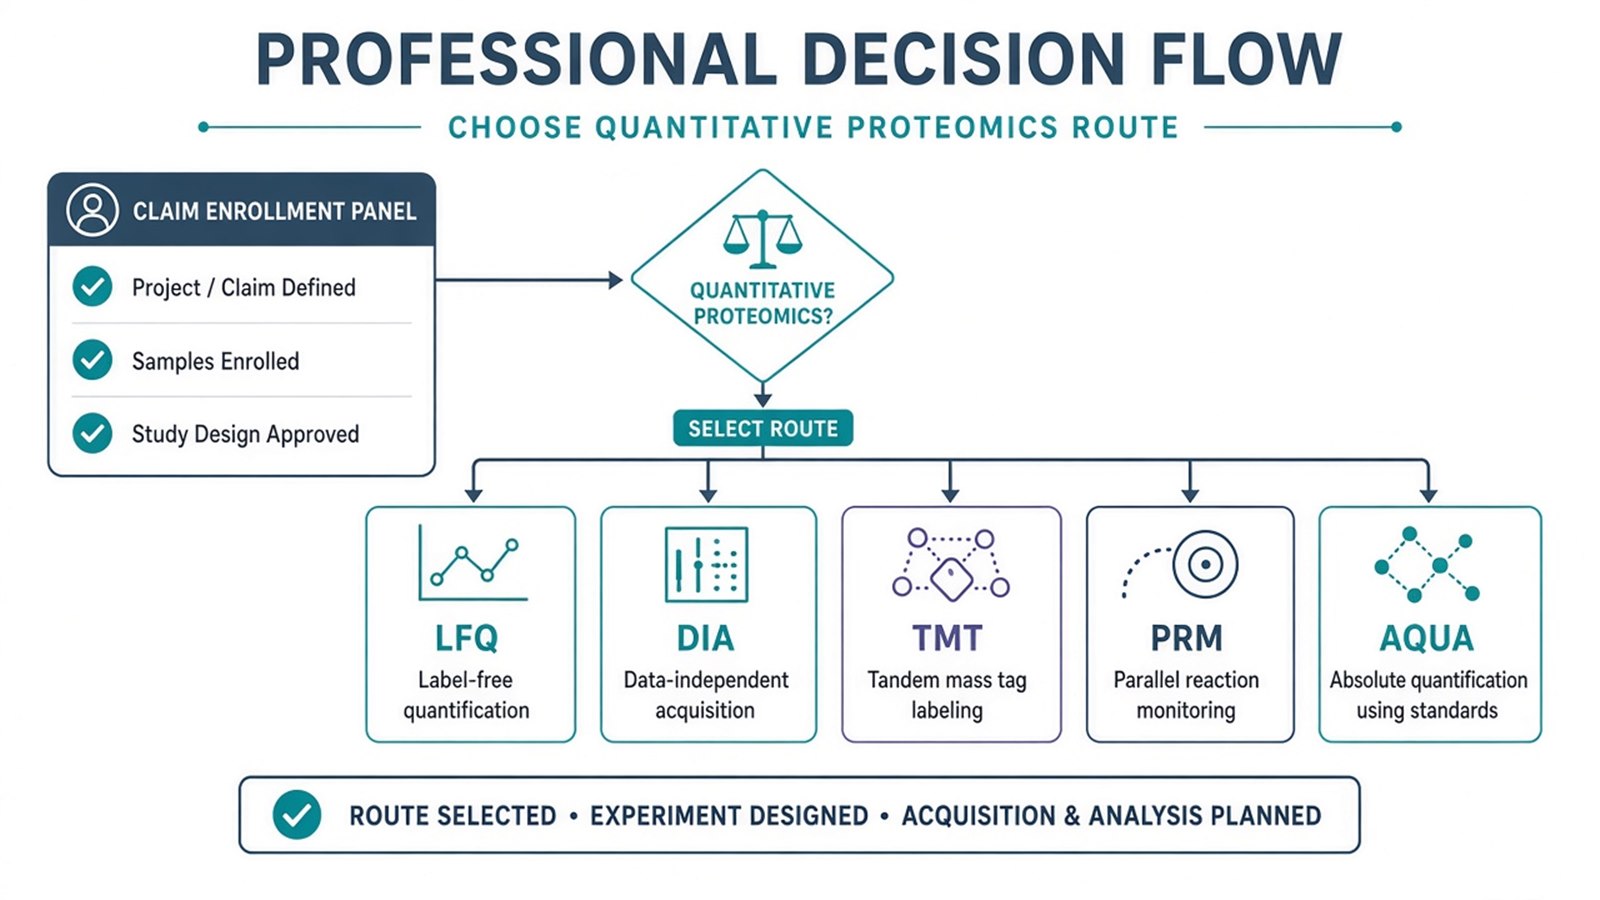
Figure 3. Method and application choice follows claim type and cohort rules, not instrument marketing alone.
For strategy comparison, see DDA vs. DIA vs. TMT in LC-MS/MS Proteomics: How to Choose the Right Strategy. For biomedical application context, see Quantitative Proteomics Analysis Empowering Biomedical Research. For prep impact, see The Impact of Sample Preparation on Quantitative Results in Quantitative Proteomics.
Limits and Application Boundaries
Quantitative proteomics does not by itself certify clinical diagnostics or drug approval claims. Relative discovery lists are not diagnostic cutoffs. Absolute claims require explicit standard designs. Batch effects, library mismatch, ratio compression, and missing values are method-specific limits, not solved by software alone. Pathway enrichment shows overrepresentation relative to the quantified background; it does not prove pathway activity.
Typical applications when methods and claims align:
- Mechanism or treatment contrasts with relative protein change.
- Biomarker-candidate discovery followed by targeted confirmation.
- Multi-arm drug or genotype panels inside a TMT plex when enrollment can freeze.
- Biofluid or tissue cohorts using label-free LFQ or DIA with flexible enrollment.
- Pharmacodynamic or biopharm monitoring on closed targeted panels with optional SIS.
Diagnostic cutoffs, companion diagnostics, and personalized treatment rules sit outside ordinary research quantitative proteomics reporting unless separately validated assay frameworks exist.
Once method family, workflow branch, and claim type are defined, a project can be scoped to quantify proteins under a defensible design. Researchers can review the MtoZ Biolabs Label-Free Quantitative Proteomics Service and TMT Quantitative Proteomics Analysis Service for sample evaluation and workflow planning.
Frequently Asked Questions
1. What does quantify protein mean in proteomics?
It means estimating relative or absolute abundance for proteins or surrogate peptides after declared prep, LC-MS/MS, and analysis rules.
2. Is label-free always simpler than labeled quantification?
Chemistry can be simpler, but independent runs require strict prep and batch control. Labeled routes add tagging but can multiplex in one pooled run.
3. When is absolute quantification required?
When the study needs amount units on a closed panel with standards and calibration, not when ordinary discovery intensities are enough.
4. Can one quantitative proteomics run serve discovery and clinical cutoff setting?
Discovery and validated cutoffs are separate designs. Discovery nominates candidates; cutoffs need targeted assay validation.
5. How do DIA and LFQ fit under quantitative proteomics?
Both are usually label-free relative routes that differ in acquisition and extraction, not in replacing targeted confirmation when needed.
Reference
- J. Cox, M.Y. Hein, C.A. Luber, et al. (2014). Accurate proteome-wide label-free quantification by delayed normalization and maximal peptide ratio extraction, termed MaxLFQ. Mol. Cell. Proteomics, 13, 2513-2526. https://doi.org/10.1074/mcp.M113.031591
- A. Thompson, J. Schafer, K. Kuhn, et al. (2003). Tandem mass tags: a novel quantification strategy for comparative analysis of complex protein mixtures by MS/MS. Anal. Chem., 75, 1895-1904. https://doi.org/10.1021/ac0262560
- A.C. Peterson, J.D. Russell, D.J. Bailey, M.S. Westphall, J.J. Coon (2012). Parallel reaction monitoring for high resolution and high mass accuracy quantitative, targeted proteomics. Mol. Cell. Proteomics, 11, 1475-1488. https://doi.org/10.1074/mcp.O112.020131
- J.R. Wisniewski, A. Zougman, N. Nagaraj, M. Mann (2009). Universal sample preparation method for proteome analysis. Nat. Methods, 6, 359-362. https://doi.org/10.1038/nmeth.1322
