Comparison of Mass Spectrometry MRM With iTRAQ/TMT Quantification
Choose MRM when a closed peptide list needs scheduled precursor-to-product transitions for relative confirmation or absolute amount with standards; choose iTRAQ or TMT when many proteins must be compared as relative ratios inside one isobaric plex after amine labeling and pooling (Lange et al. 2008; Thompson et al. 2003). MRM is targeted acquisition on typically a triple quadrupole; iTRAQ/TMT are multiplex labeling chemistries usually read by discovery-style LC-MS/MS reporters. They answer different claims and do not replace each other: MRM does not survey an open proteome, and ordinary TMT or iTRAQ reporters are not absolute concentration. Prefer discovery LFQ or DIA first when candidates are still unknown.
Researchers who already have defined samples, comparison groups, or a specific project objective can also review the MtoZ Biolabs SRM-MRM Quantitative Proteomics Services and TMT Quantitative Proteomics Analysis Service for project-specific feasibility and assay planning.
Comparison at a Glance: Different Method Layers
Dimension |
MRM (also called SRM) |
iTRAQ / TMT |
|---|---|---|
Method layer |
Targeted acquisition schedule |
Isobaric labeling chemistry |
What is measured |
Predefined peptide transitions |
Reporter ions from pooled labeled channels |
Inventory |
Closed peptide list required |
Open discovery inside one plex is common |
Enrollment |
Independent injections of the panel |
Must freeze samples before pooling |
Default claim |
Relative confirmation; absolute only with SIS |
Relative ratios inside the plex |
Typical strength |
Selectivity for nominated targets |
Same-run multi-sample multiplex discovery |
Typical limit |
Does not find unscheduled proteins |
Ratio compression; no late enrollment |
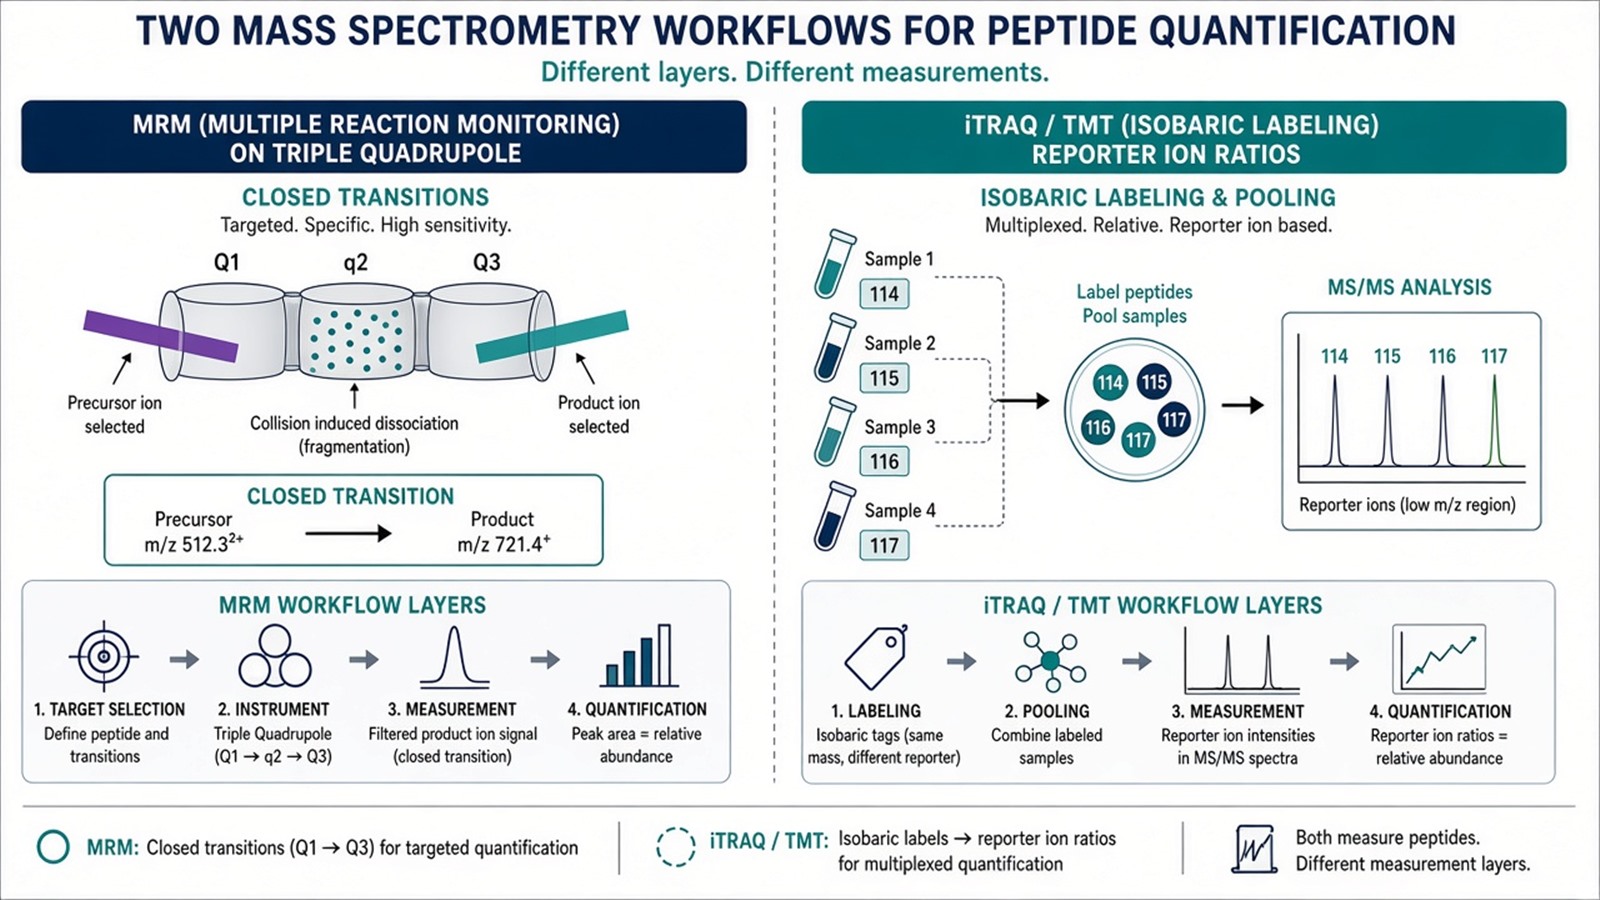
Figure 1. MRM and iTRAQ/TMT are not interchangeable quantification recipes: one is targeted acquisition; the other is multiplex isobaric labeling.
Ranking “sensitivity” without stating panel status, plex closure, and relative-versus-absolute claim misleads method selection.
How to Choose: Selection Criteria
Selection question |
Prefer MRM |
Prefer iTRAQ / TMT |
Prefer another route |
|---|---|---|---|
Are proteotypic peptides already nominated? |
Yes |
Not required for discovery inside the plex |
No → LFQ or DIA first |
Must many conditions share one MS run? |
Not the main reason |
Yes, if amine chemistry and loads match |
Enrollment open → LFQ/DIA |
Is absolute peptide amount required? |
Yes, with SIS and calibration |
No by default |
AQUA/PRM designs may fit better |
Is the matrix complex and the panel still refining? |
Consider PRM instead or after MRM |
Discovery plex may still help nominate |
Unit-resolution interference dominates |
Is sample input too scarce for tags and QC? |
Possible if panel is tiny |
Often blocked |
Unlabeled discovery |
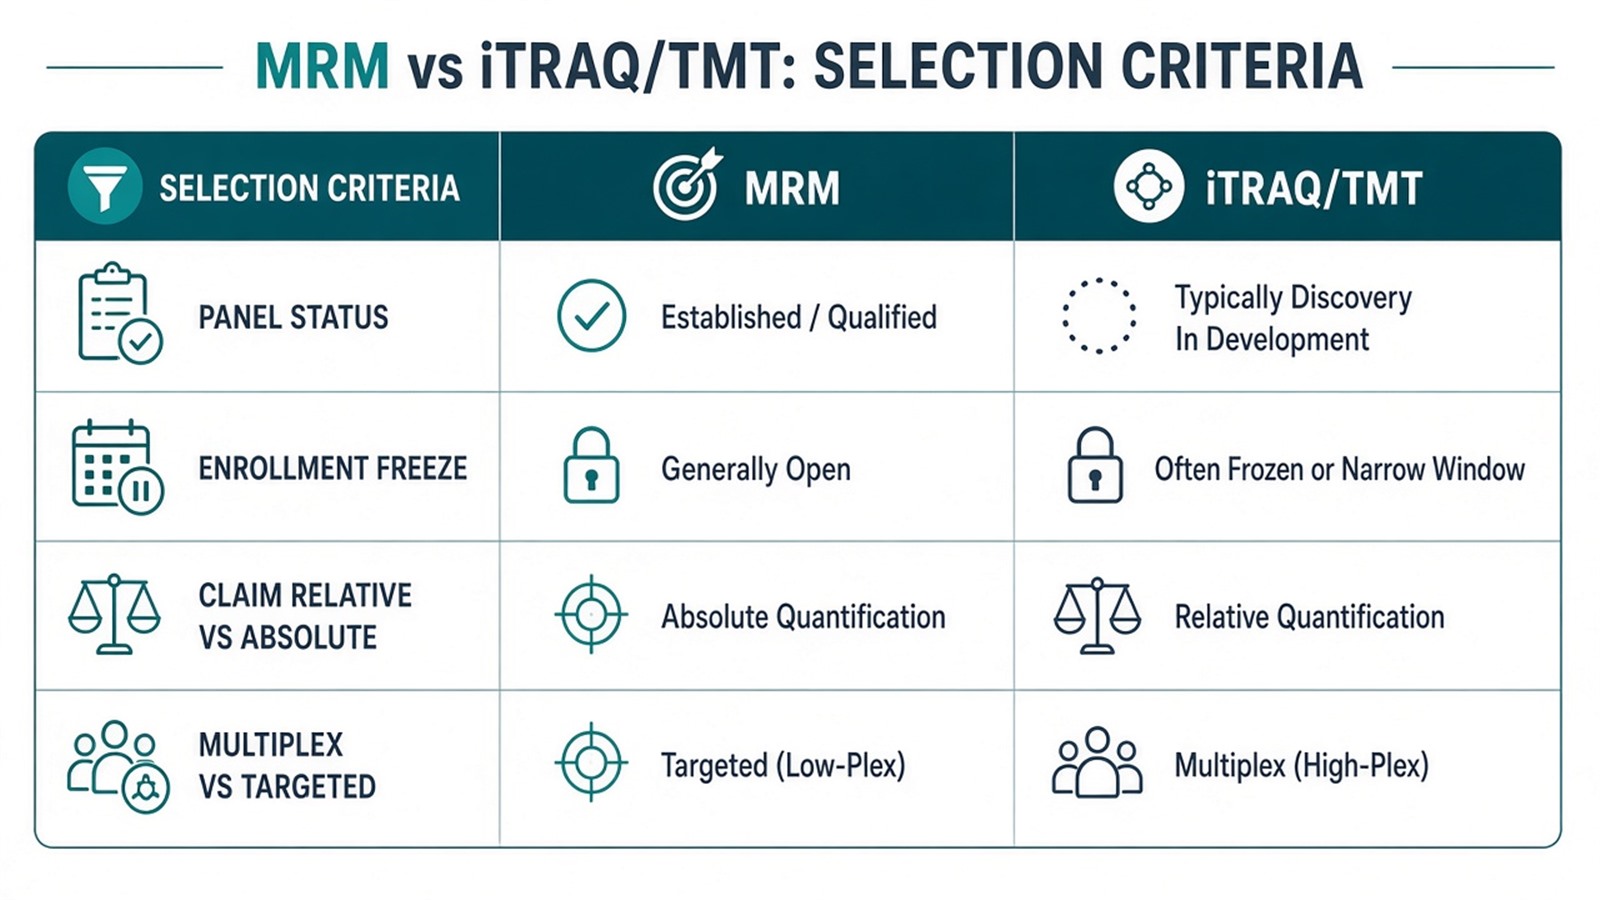
Figure 2. Selection follows claim and design gates: closed-list targeting versus frozen multiplex labeling.
iTRAQ and TMT share the isobaric-label family. Kit chemistry and channel count differ, but both require matched amine-compatible prep and a frozen map. MRM shares the closed-list idea with PRM; PRM stores full high-resolution MS2 and often fits complex digests during panel refinement (Peterson et al. 2012).
For TMT principle and workflow, see TMT Protein Quantification and Tandem Mass Tag Quantitative Proteomics. For MRM principle, see Principle of Multiple Reaction Monitoring and SRM Mass Spectrometry. For PRM versus MRM, see PRM vs MRM: Comparative Analysis in Quantitative Proteomics. For validation planning after discovery, see How to Plan a Targeted Proteomics Validation Study.
Judgment Criteria: When Each Route Fits
Study need |
MRM often fits |
iTRAQ / TMT often fit |
Reassess |
|---|---|---|---|
Claim |
Confirm named peptides; optional absolute with SIS |
Relative multi-arm discovery in one plex |
Diagnostic cutoff without validated framework |
Scale |
Short to moderate closed panel |
Broad relative proteome inside kit capacity |
Panel unknown and plex cannot freeze |
Chemistry |
No isobaric labeling required |
Amine cleanup and equal load feasible |
Residual amines or scarce input |
Sequence in a program |
After LFQ/DIA/TMT nominates candidates |
Early multi-condition screen when plex fits |
Treating MRM as open discovery |
Prefer a common research sequence when needed: unlabeled or isobaric discovery → close a shortlist → MRM or PRM confirmation → add SIS only if amount units are required. That sequence is a project plan, not a rule every study must follow.
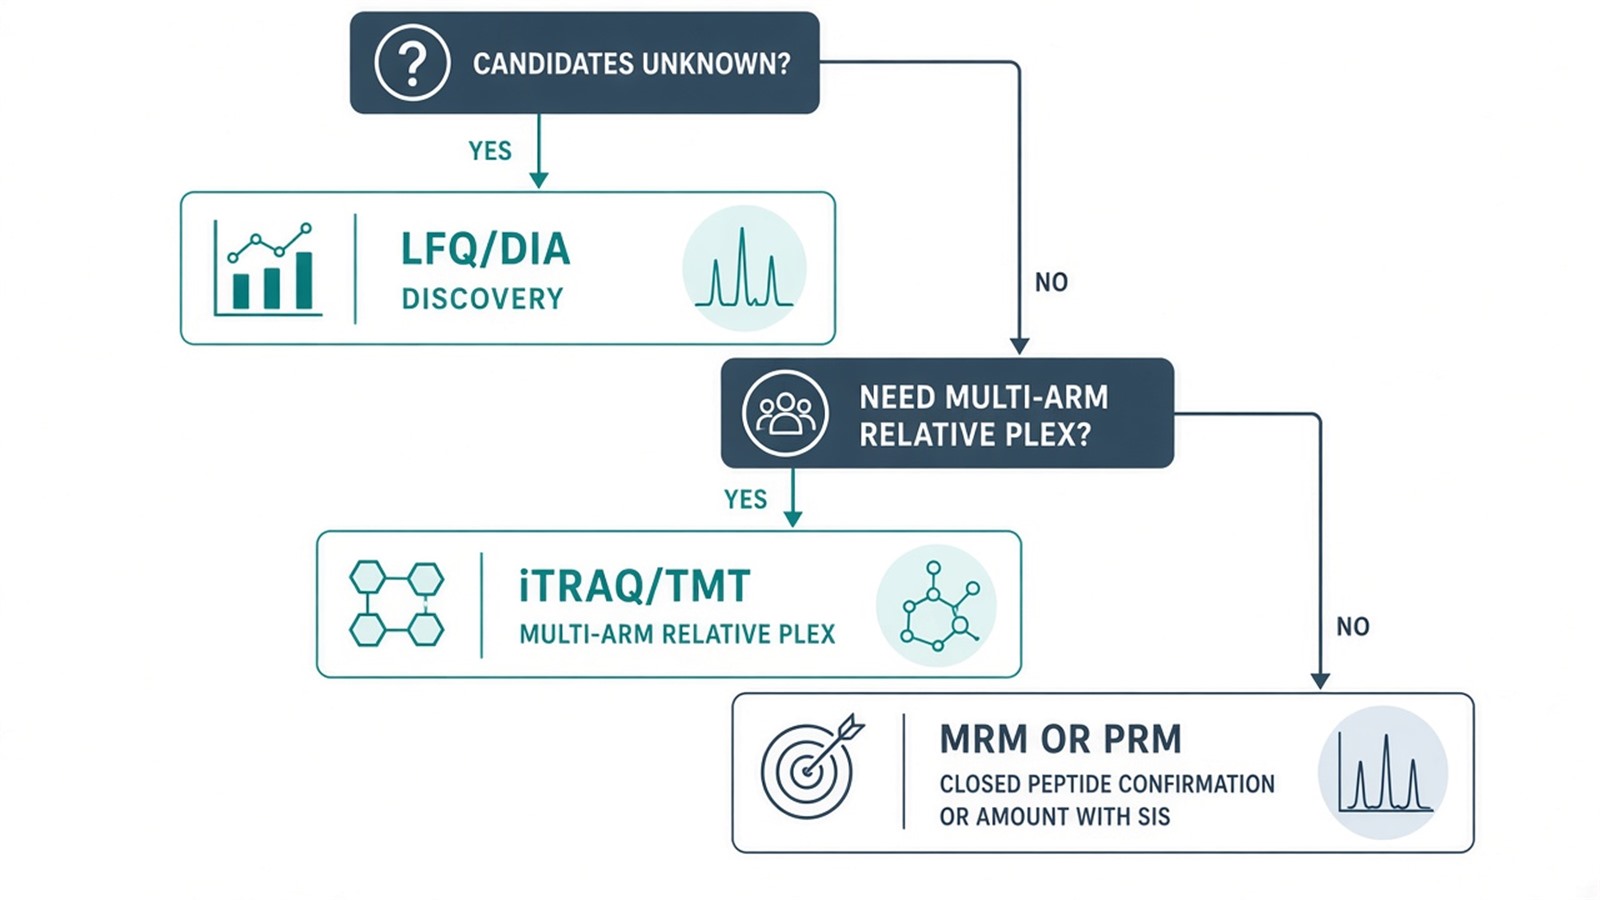
Figure 3. Choose MRM for closed-list targeting and iTRAQ/TMT for frozen multiplex relative discovery; keep LFQ or DIA when candidates are still unknown.
For labeling versus unlabeled discovery, see Comparing LFQ and Isobaric Labeling Strategies in Proteomics Quantification. For absolute designs on MRM or PRM, see Absolute Quantification in Targeted Proteomics and AQUA Proteomics.
Limits and Application Boundaries
MRM does not measure peptides absent from the transition list. iTRAQ/TMT do not accept late samples after pooling and do not create absolute concentration from reporters alone. Ratio compression can shrink TMT or iTRAQ fold changes; unit-resolution MRM can suffer transition interference. Pathway enrichment from either route reflects the quantified background of that experiment, not proven pathway activity.
Typical paired applications:
- Multi-arm drug or genotype screens by TMT or iTRAQ, then MRM confirmation of a shortlist.
- Biomarker-candidate validation by MRM after discovery nominates proteotypic peptides.
- Pharmacodynamic closed panels by MRM when enrollment stays open across many independent injections.
- Absolute peptide amount by MRM plus SIS when calibrators and spike timing are planned.
Diagnostic cutoffs and companion-diagnostic claims sit outside ordinary research MRM or isobaric reporting unless a separately validated assay framework is in place.
Once panel status, plex feasibility, and claim type are defined, the project can be scoped as MRM targeting, iTRAQ/TMT multiplex discovery, or a staged combination. Researchers can review the MtoZ Biolabs Targeted Proteomics Service and TMT Quantitative Proteomics Analysis Service for sample evaluation and workflow planning.
Frequently Asked Questions
1. Is MRM simply more accurate than iTRAQ/TMT?
No. Accuracy follows design and QC. MRM is selective for closed targets; iTRAQ/TMT excel at multiplex relative discovery inside a plex.
2. Can iTRAQ/TMT replace MRM for absolute quantification?
Not by reporter ions alone. Absolute amount needs matched standards and calibration, commonly on a targeted platform.
3. Can MRM replace TMT for a 16-sample treatment screen?
Usually not as the sole discovery route. MRM needs a closed list; TMT can compare many proteins across channels in one mix when chemistry holds.
4. Should I choose PRM instead of MRM in this comparison?
Choose PRM when accurate-mass fragment review helps in complex digests or the panel is still refining. Choose MRM when transitions are locked and high multiplex on a triple quadrupole is required.
5. What if candidates are still unknown?
Start with LFQ, DIA, or an isobaric discovery plex. Move to MRM only after peptides are nominated.
Reference
- V. Lange, P. Picotti, B. Domon, R. Aebersold (2008). Selected reaction monitoring for quantitative proteomics: a tutorial. Mol. Syst. Biol., 4, 222. https://doi.org/10.1038/msb.2008.61
- A. Thompson, J. Schafer, K. Kuhn, et al. (2003). Tandem mass tags: a novel quantification strategy for comparative analysis of complex protein mixtures by MS/MS. Anal. Chem., 75, 1895-1904. https://doi.org/10.1021/ac0262560
- A.C. Peterson, J.D. Russell, D.J. Bailey, M.S. Westphall, J.J. Coon (2012). Parallel reaction monitoring for high resolution and high mass accuracy quantitative, targeted proteomics. Mol. Cell. Proteomics, 11, 1475-1488. https://doi.org/10.1074/mcp.O112.020131
Support Documents
Quantitative Proteomics: Label-Based and Label-Free Strategies
Compare label-based and label-free quantitative proteomics strategies and explore their applications to help identify a suitable approach for your research.
Protein Quantification for Deeper Discovery
Gain a clearer understanding of protein quantification approaches and their applications in studying protein abundance changes by downloading our detailed brochure.